Working with Enthalpies of Formation in Practice
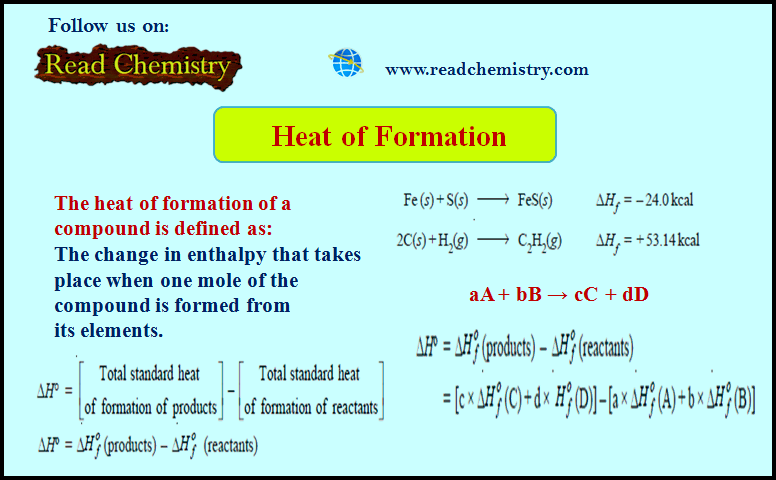
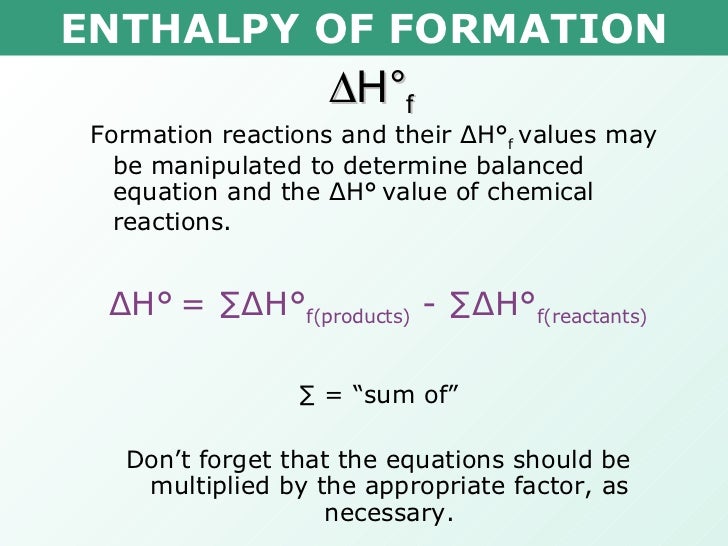
The standard enthalpy change for any reaction is calculated by subtracting the sum of formation enthalpies for your reactants from the sum for your products. That is the basic idea. The formula is straightforward: H°rxn = nH°f(products) mH°f(reactants). You plug in the values, multiply by stoichiometric coefficients, and you get your answer. I learned this in undergrad and used it constantly through my first few years in process engineering. It seems like the easiest calculation in thermodynamics because the math itself is just addition and subtraction. The problem is that most people skip the details around it and end up with answers that are wrong in ways they can't explain. Let me walk through how I actually use this. Say you're calculating the enthalpy of combustion for methane. You look up H°f values from a reliable source like the NIST Chemistry WebBook or Perry's Handbook. Methane is 74.8 kJ/mol, CO2 is 393.5 kJ/mol, and liquid water is 285.8 kJ/mol. The reaction produces one mole of CO2 and two moles of H2O, so you multiply accordingly: [1(393.5) + 2(285.8)] [1(74.8) + 2(0)]. The oxygen term drops out because it's an element in its standard state, which brings me to something that trips people up regularly.
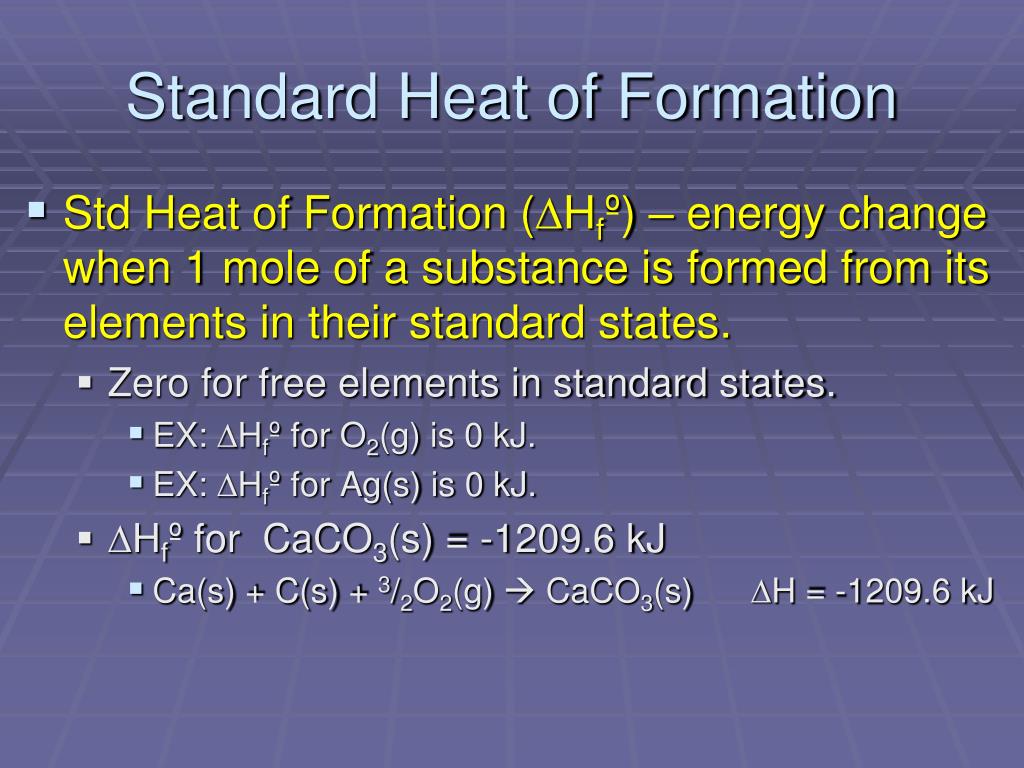
Elements in their standard states have H°f = 0 by definition. This applies to O2(g), N2(g), H2(g), C(s, graphite), Br2(l), I2(s), and a few others. The graphite specification matters. If you see a table listing diamond with a non-zero value, that's correct because diamond is not the standard state. I've seen students use the diamond value for carbon and wonder why their answer was off by about 2 kJ/mol. Here's the edge case that cost me an afternoon once. I was working on a heat balance for a reactor running at elevated temperature, and I needed to adjust standard enthalpies to process conditions. I pulled H°f values from a database and tried to use them directly in a Kirchoff's law integration. The numbers didn't close. I spent about two hours debugging before I realized the database had listed water as a gas at 241.8 kJ/mol while my reference data had it as a liquid at 285.8 kJ/mol. That's a 44 kJ/mol difference, which is exactly the enthalpy of vaporization. For reactions producing water, using the wrong phase value skews your result significantly. I switched everything to gas-phase values since the reactor effluent was above the dew point, and the balance closed. The takeaway is simple: always verify the phase and temperature reference for every value you pull. Another thing that doesn't get enough attention is how these values are actually determined. Most H°f values come from bomb calorimetry for combustion reactions. The calorimeter measures the heat released when a substance burns completely, and then you back-calculate the formation enthalpy from the known products. This means compounds that are difficult to burn cleanly or that decompose before burning will have less reliable values. Hydrogen bonding compounds, large organic molecules with multiple functional groups, and certain intermediates fall into this category. Their tabulated values might carry uncertainty in the ±2 to ±5 kJ/mol range, sometimes more.
When you're dealing with a reaction network rather than a single equation, the approach scales but gets messier. You still apply the same summation across all species, but you need to be careful about the stoichiometric coefficients matching your balanced equation exactly. I once reviewed a calculation where someone used fractional coefficients for a catalyst-mediated pathway and forgot that the catalyst shouldn't appear in the enthalpy sum at all. It added a small error, but in a sensitivity analysis it propagated through the whole model. For reactions far from standard conditions, the Heat Of Formation Equation gives you the baseline at 298.15 K and 1 bar. If your system operates at 500 K or higher, you need to account for temperature dependence. The standard approach uses heat capacity correlations. You integrate Cp dT from 298 K to your target temperature for each species, then apply those corrections to the formation enthalpies before running the product-minus-reactant calculation. Polynomial Cp expressions from databases like JANAF or the NIST-JANAF Thermochemical Tables work well here. Third-order polynomials in T are common, though some sources use Shomate equations which are equivalent but expressed differently. Phase changes complicate this too. If a component crosses its boiling or melting point between 298 K and your operating temperature, you can't just integrate Cp across the transition. You need to add the latent heat term at the appropriate temperature. A single smooth Cp polynomial won't capture that discontinuity. I've seen this overlooked in quick hand calculations where the error ended up being larger than the rounding from other approximations.
Get the Full Details
The biggest limitation of relying on standard formation enthalpies is that they assume ideal behavior at standard pressure. In real industrial systems, especially at pressures above 10 bar or with highly non-ideal mixtures, fugacity corrections matter. You'd move to a real gas equation of state or activity coefficient model. The formation enthalpy values themselves don't change much with pressure for condensed phases, but for gases the enthalpy does shift slightly. At moderate pressures the shift is small, but it's not zero and it accumulates in multi-stage calculations. If you need this for computational work, most process simulation packages have built-in property methods that handle the temperature and pressure corrections automatically. Aspen Plus, HYSYS, and similar tools use databanks derived from the same standard sources I mentioned, but they add the necessary corrections internally. The manual approach is still useful when you're doing a quick sanity check or working outside a simulator. It also helps you catch when the software is using a default property method that doesn't match your system's chemistry. One practical tip that saves time: keep a short list of the most commonly needed H°f values memorized or in a quick-reference sheet. CO2, H2O(l), H2O(g), NH3, NO, NO2, SO2, and a few others come up constantly. Having them at hand lets you do rough calculations in minutes instead of spending time hunting through tables. The rough calculations are often good enough to spot order-of-magnitude errors before you commit to a detailed model.