The Math Behind Meiotic Shuffling
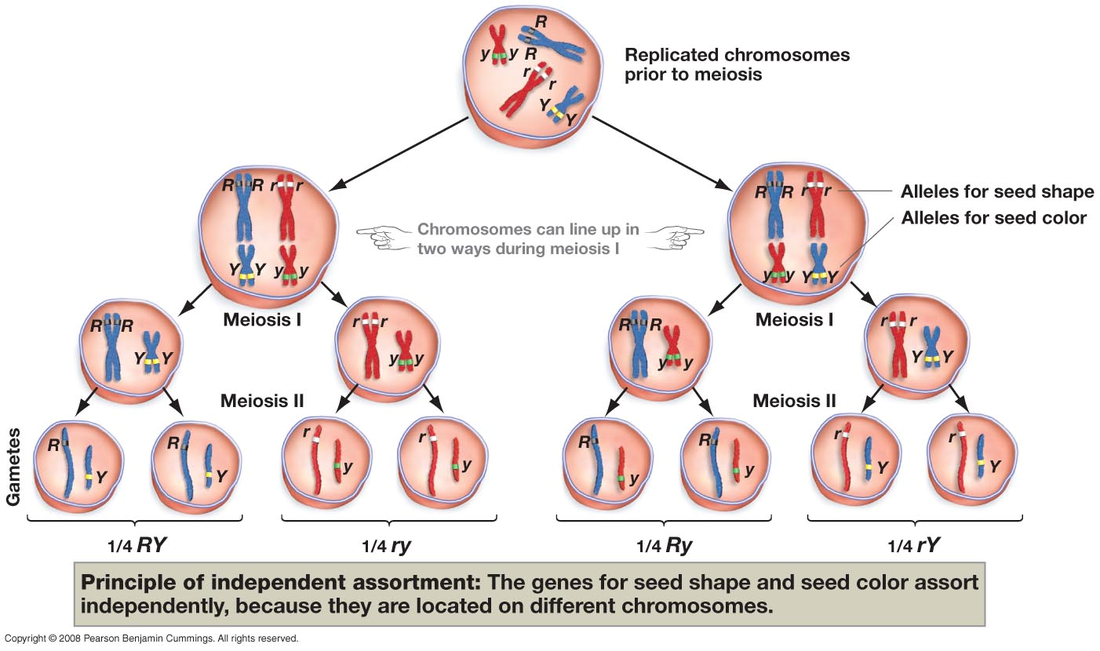
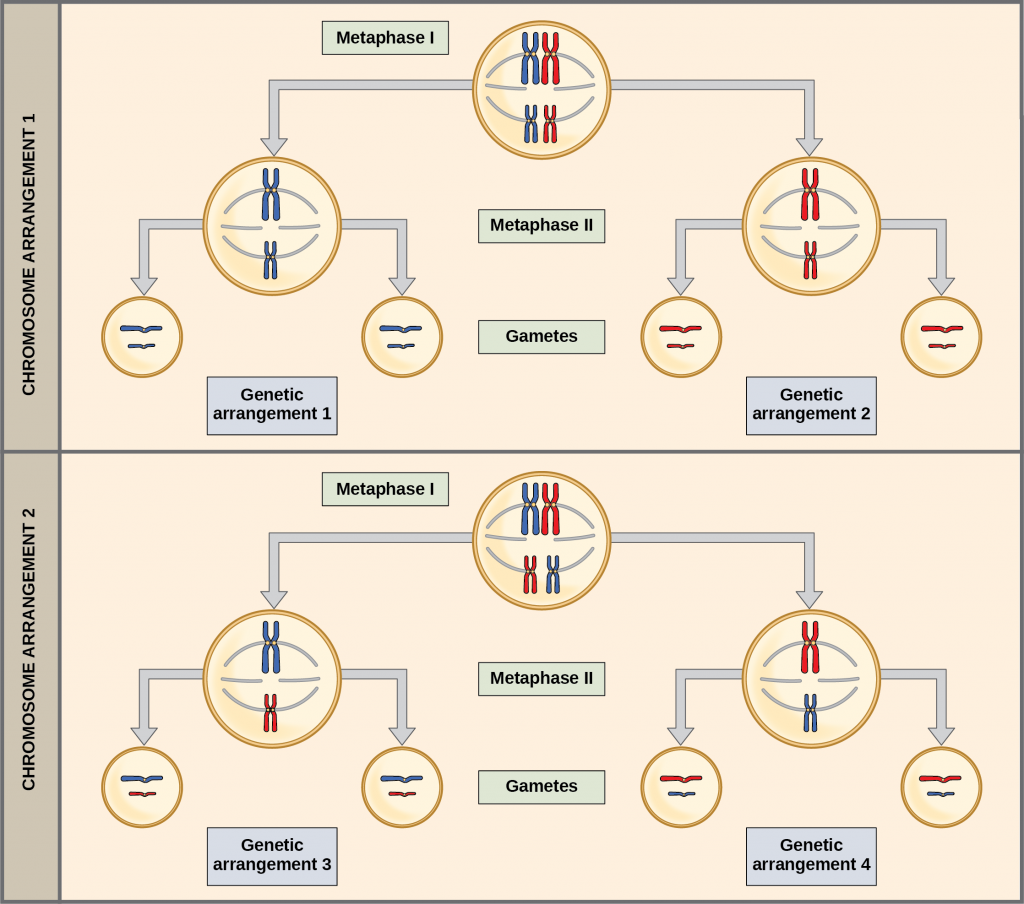
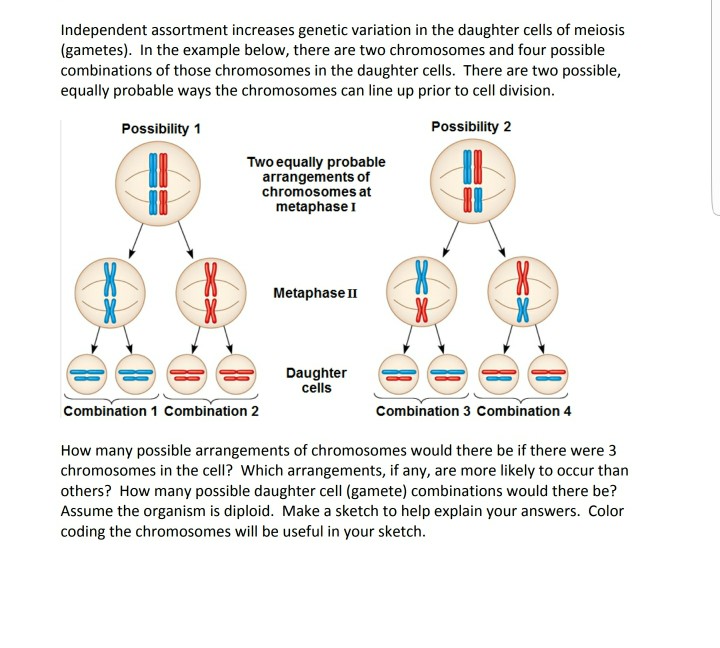
The short answer is that the number of unique gametes from independent assortment equals 2 to the power of n, where n is the number of heterozygous gene pairs. That is the textbook formula. It works cleanly in practice when you have the right assumptions. It falls apart quickly if you ignore real biological constraints. Let me show you how this actually plays out. Say you have an organism that is AaBbCcDd at four independent loci. Each heterozygous pair segregates independently during metaphase I. That gives you 2 to the 4th power, which is 16 possible gamete genotypes. Each one occurs at roughly equal frequency if everything behaves normally. Now multiply that out for three loci and you are already at 8. Six loci gets you to 64. Ten loci jumps to 1024. Twelve hits 4096. The numbers grow fast and they do not care about your comfort level.
How Many Unique Gametes Could Be Produced Through Independent Assortment
The calculation itself is straightforward arithmetic once you count correctly. The hard part is counting correctly. I have spent more time correcting students' work on this than I have actually teaching the formula. The most common error is including homozygous loci in n. If a locus is AA or aa, it contributes zero to the diversity. It only produces one type of allele. Do not double count it. Here is a practical example from a genetics lab I ran a few years back. I was working with a set of Drosophila stocks that had multiple marked balancer chromosomes and a bunch of unmarked segregating loci in the background. The theoretical calculation suggested around 512 distinct gamete types across ten heterozygous sites. In practice, after scoring progeny over three generations, I recovered only about 380 visibly distinct genotype combinations. The missing ones were not a math problem. They were a viability problem. Several recombinant classes carried unfavorable allele combinations that reduced larval survival. The gametes were produced. They just did not make it to adulthood. That is the first nuance people miss. The 2^n formula tells you about gamete genotypes, not about viable offspring. Two things can differ. Chromosome number matters enormously. Humans have 23 pairs, so the theoretical maximum is 2 to the 23rd, which equals about 8.4 million unique gamete combinations per individual. Eight point four million. That is without crossing over. Once you add recombination into the mix, the number becomes astronomically larger and completely uncountable by hand.
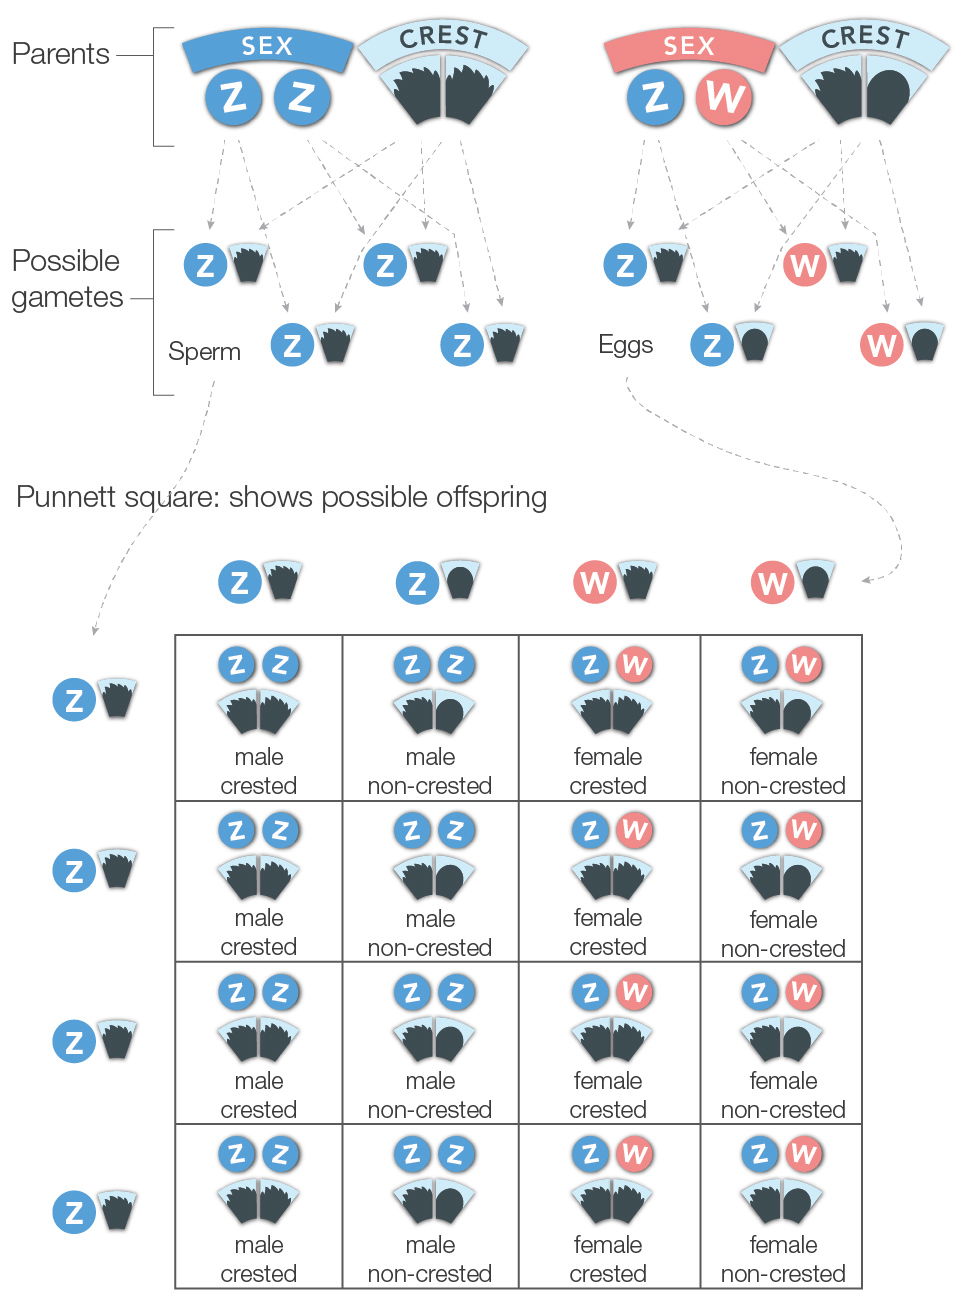
I worked through a problem once with a graduate student who tried to enumerate all possible gametes for a wheat line with hexaploid genetics. Hexaploid. Three copies of each chromosome. The standard 2^n formula does not apply at all. You have to think about multinomial segregation. I spent two weeks building a small Python script that simulated chromosome segregation under different pairing configurations. The script showed that even for a modest number of heterozygous loci, the combinatorial space exceeded practical enumeration. We switched to estimating effective gamete diversity using allele frequencies and Hardy-Weinberg approximations instead. It gave us a usable number in about fifteen minutes. The full enumeration would have required more compute time than we had budgeted for the project. Another edge case I keep running into involves linked genes. Independent assortment only applies to genes on different chromosomes or genes far enough apart on the same chromosome that they behave as if unlinked. The rule of thumb is roughly 50 centimorgans or more. Below that, you get linkage disequilibrium and the simple formula overestimates diversity. I had a mapping project where I assumed six heterozygous loci segregated independently and calculated 64 gamete classes. The testcross data showed only 36 observable classes with strong bias toward parental types. The loci were clustered within a 20 centimorgan window. I ended up calculating expected gamete frequencies using recombination fractions instead. Much less elegant. More accurate. There is also the issue of sex determination systems. The 2^n formula assumes autosomal loci. If you include the sex chromosomes, the math changes. A human male is XY. The X and Y do not assort in the same way as autosomes. Sperm carry either an X or a Y, but the gene content differs. For a male who is heterozygous at autosomal loci, you calculate 2^n for the autosomes and then factor in the sex chromosome separately. It is a small adjustment. It matters when you are doing precise pedigree analysis or calculating inheritance probabilities for X-linked traits.
Get the Full Details

Sister chromatid cohesion and nondisjunction events also break the model. If chromosomes fail to separate properly during meiosis I or II, you get aneuploid gametes. The formula assumes perfect segregation. It does not. In human oocytes, nondisjunction rates increase with maternal age. A 35-year-old woman has a measurably higher rate of trisomic conceptions than a 25-year-old. The underlying gamete production mechanism is the same. The error rate is not. This means the theoretical number of unique gametes is always an upper bound. The actual number reaching fertilization is lower. If you need to calculate this for a specific organism, here is the method I use. First, identify every locus where the individual is heterozygous. Count them. Call that number n. Calculate 2^n. Then check for linked loci within the same chromosome. If any heterozygous loci are closer than about 20 centimorgans, adjust your expectation downward based on the observed recombination fraction. If the organism is polyploid, switch to a different model entirely. If you are working with data rather than predictions, generate a Punnett square or use a gamete frequency table only if n is small. Beyond n equals about 6, you are better off using a computational tool. I recommend a simple script over manual enumeration. Here is what I use when I need to be precise. It takes a list of heterozygous loci and outputs all possible gamete combinations:
import itertools

genotypes = ['Aa', 'Bb', 'Cc', 'Dd']
alleles = [list(itertools.product(g.replace('A','A').replace('a','a'), repeat=1)) for g in genotypes]

Simpler approach:
loci = ['Aa', 'Bb', 'Cc']
gametes = [''.join(x) for x in itertools.product(*[[loc[:1], loc[1:]] for loc in loci])]
print(f'Number of unique gametes: {len(gametes)}')
for g in gametes:
    print(g)
This prints 8 gametes for three heterozygous loci. Change the input list and it scales. I run this before any manual work because it catches errors in my counting. I once thought I had five heterozygous loci in a plant cross and calculated 32 gamete types. The script showed six. I had misread a gel image. The extra band was homopolymer slippage, not a real allele. The script would have saved me three hours of recalculating pedigree ratios. There is also a conceptual trap around what counts as unique. Genotypically unique and phenotypically unique are different questions. If two gamete genotypes produce the same phenotype due to dominance or epistasis, they are not functionally different in a breeding program. I once designed a crossing scheme based on genotypic diversity and was confused when the phenotypic variance in the F2 generation was much lower than expected. Seven of the 64 theoretical gamete combinations collapsed into three observable phenotypes because of complete dominance at two loci. The genetics was correct. The interpretation was not. For most classroom problems, the answer is 2^n with n equal to heterozygous pairs. For real research, the answer depends on linkage, ploidy, segregation distortion, and viability. The formula is a starting point, not a finish line. I have seen people treat it as gospel and then get burned when empirical data did not match. The data is always right. The model is just a model.
If you are working with a specific organism and need help figuring out whether your loci are truly assorting independently, share the map distances or chromosome locations. That tells you more than the heterozygous count alone. Raw numbers without genomic context will mislead you every time.