Getting the Numbers Without Losing Your Mind
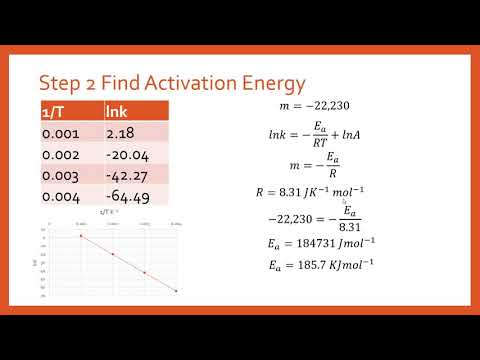
Activation energy is the energy barrier a reaction has to clear before it actually proceeds. You find it experimentally by measuring reaction rates at different temperatures and feeding those numbers into the Arrhenius equation. The equation itself is k = A * exp(-Ea/RT), where k is the rate constant, A is the pre-exponential factor, Ea is the activation energy, R is the gas constant, and T is temperature in Kelvin. The practical way most people actually do this in a lab or a data analysis project is by linearizing the equation. Take the natural log of both sides and you get ln(k) = ln(A) - Ea/(R*T). Plot ln(k) on the y-axis against 1/T on the x-axis. The slope of that line is -Ea/R. Multiply the slope by -R and you have your activation energy in joules per mole.
How To Calculate Activation Energy Using the Two-Point Form
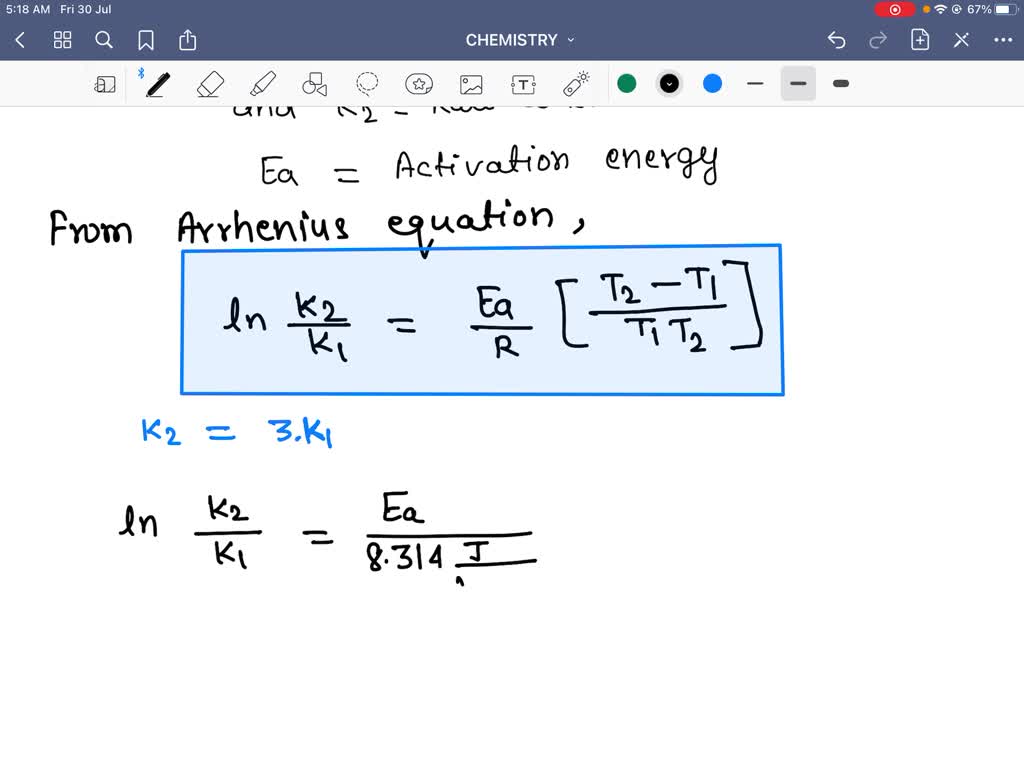
If you only have two data points instead of a full dataset, you can skip the regression and use the two-point version of the Arrhenius equation directly: ln(k2/k1) = (Ea/R) * (1/T1 - 1/T2). Rearrange to solve for Ea: Ea = R * ln(k2/k1) / (1/T1 - 1/T2). It gives the same answer that a linear fit would give through those two points. That is useful when you are just checking something quickly and do not need high precision. The bigger question is usually where your rate constants come from. In a real kinetics experiment you measure concentration versus time, extract a rate constant for each temperature, and then run the calculation. If you are working with first-order reactions the math is straightforward. You take the natural log of concentration over time and the slope gives you -k. Second-order and more complex rate laws require a bit more care with the integrated forms, but the principle stays the same. I have seen people skip the extraction step and just use half-lives or reaction times as proxies for k. That works only if every measurement was made under identical initial conditions and the reaction order does not change across temperatures. When the mechanism shifts even slightly at higher temperatures, your proxy introduces systematic error that the Arrhenius plot will faithfully amplify.
What Actually Goes Wrong in Practice
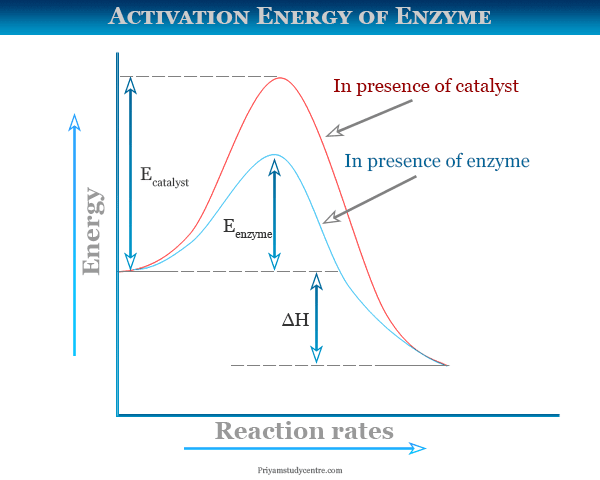
The most common mistake is forgetting to convert Celsius to Kelvin. It sounds ridiculous until you see someone get an activation energy that is roughly three times too large because they used 25, 35, and 45 as their temperatures. The other typical error is mixing units for R. If your answer needs to be in kilojoules per mole, use R = 8.314 J/(mol*K) and divide the final result by 1000. Using the calorie-based value of R without adjusting leads to numbers that look plausible but are completely wrong. Temperature control matters more than most people expect. A drift of even one degree Celsius at around 300 Kelvin changes 1/T enough to noticeably shift the slope. I worked on a project a few years ago measuring the degradation rate of a pharmaceutical intermediate across five temperatures. The thermostat on the water bath was calibrated wrong by about 1.2 degrees. Our initial Arrhenius plot showed a beautiful straight line with an R-squared of 0.998, which should have been a red flag right there. Real data rarely looks that clean. We recalibrated with a NIST-traceable thermometer, repeated the runs, and the slope shifted by about eight percent. The activation energy dropped from roughly 82 kilojoules per mole to about 75. That eight percent difference is the kind of thing that matters when you are making stability predictions for a product shelf life. Another issue that comes up constantly is the assumption that the activation energy is constant across your temperature range. It is not always. Reactions with competing pathways, enzyme-catalyzed systems, or reactions where the rate-determining step changes with temperature will produce curved Arrhenius plots. A curve is not a failure of the math. It is information. If your plot bends, you either have multiple mechanisms active or your temperature range is too wide for a single Ea to describe. Split the data into separate ranges and fit each segment individually, or switch to a more appropriate model.
Get the Full Details
The Spreadsheet or Script Workflow
Most people end up doing this in a spreadsheet or a quick Python script. Here is the straightforward path. Put your temperatures in one column and convert them to Kelvin. Add another column for the reciprocal of temperature. Put your rate constants in a third column and a fourth column for the natural log of those constants. Use a linear regression on the ln(k) versus 1/T columns. The slope times negative R gives you Ea. If you are using Python, numpy and scipy make this trivial. Fit a line with np.polyfit on the transformed data or use scipy.optimize.curve_fit for a direct nonlinear fit to the exponential form. The nonlinear fit is technically more statistically sound because it does not assume the residuals are normally distributed in log space, but for typical laboratory data with reasonable measurement precision the linear and nonlinear results will be essentially identical. For people who prefer not to code, I routinely hand out a simple Google Sheets template that handles the conversions and regression automatically. You paste in temperature and rate data, and the sheet outputs Ea, the pre-exponential factor, and the standard errors. The link is below.
Download the Arrhenius Calculator Sheet
When This Method Completely Falls Apart
The Arrhenius approach assumes a single elementary step or a dominant pathway with a fixed energy barrier. Diffusion-controlled reactions in solution often show much weaker temperature dependence because the viscosity of the solvent changes with temperature in a way that masks the true activation barrier. Photochemical reactions are another area where the concept of activation energy becomes awkward. The energy input comes from photons, not thermal collisions, so an Arrhenius analysis may produce numbers that are internally consistent but chemically meaningless. Heterogeneous catalysis introduces another layer of complexity. Adsorption equilibria shift with temperature, and what you are really measuring is a composite of the activation energy for surface reaction and the temperature dependence of the adsorption term. The Eyring equation from transition state theory can sometimes untangle this better than the Arrhenius form, but it requires knowing or estimating the entropy of activation, which is not always available. There is also the issue of what R you actually use in different unit systems. If your rate constants are in per second and your activation energy needs to come out in kilocalories per mole, you have to use R = 1.987 cal/(mol*K) and convert at the end. I lose count of how many times I have caught a student using 8.314 and getting an answer in joules and not realizing it because the number looked reasonable.

A Few Practical Details People Overlook
Error propagation matters. The uncertainty in Ea depends on the spread of your temperature points and the scatter in your rate constants. Wider temperature ranges give tighter confidence intervals on the slope. If all your measurements fall within a ten-degree window, your activation energy will come with a fairly wide error bar even if your rate constant measurements are precise. I aim for at least forty to fifty degrees of span whenever possible. The pre-exponential factor A is not just a fitting parameter. It carries physical meaning related to the frequency of effective collisions or the attempt frequency in transition state theory. Extracting a reasonable A value alongside Ea is a useful sanity check. If your A comes out to something like 10 to the minus five per second for a reaction that should be diffusion-limited, something is wrong with the data or the assumed reaction order. Typical A values for unimolecular reactions sit around 10 to the 13 per second, and for bimolecular reactions in solution they are often in the 10 to the 10 to 10 to the 12 range depending on the units. Nonsensical A values are a faster diagnostic than any statistical test on the regression. One more thing that causes problems is using initial rate data without confirming that the reaction remains in the initial rate regime at every temperature. As temperature increases, side reactions and product inhibition can kick in faster than the main reaction, and your extracted k will be systematically biased. Running the reaction for a shorter time at higher temperatures and verifying linearity of the early concentration data prevents this.
For most routine work, the linearized Arrhenius method with three or more temperature points gives a solid activation energy in about fifteen minutes once your data is in hand. The method breaks down predictably when the underlying assumptions are violated, so checking the plot for curvature and the A value for physical reasonableness catches the vast majority of problems before they propagate into published numbers.