Understanding Enthalpy Calculations in Practice
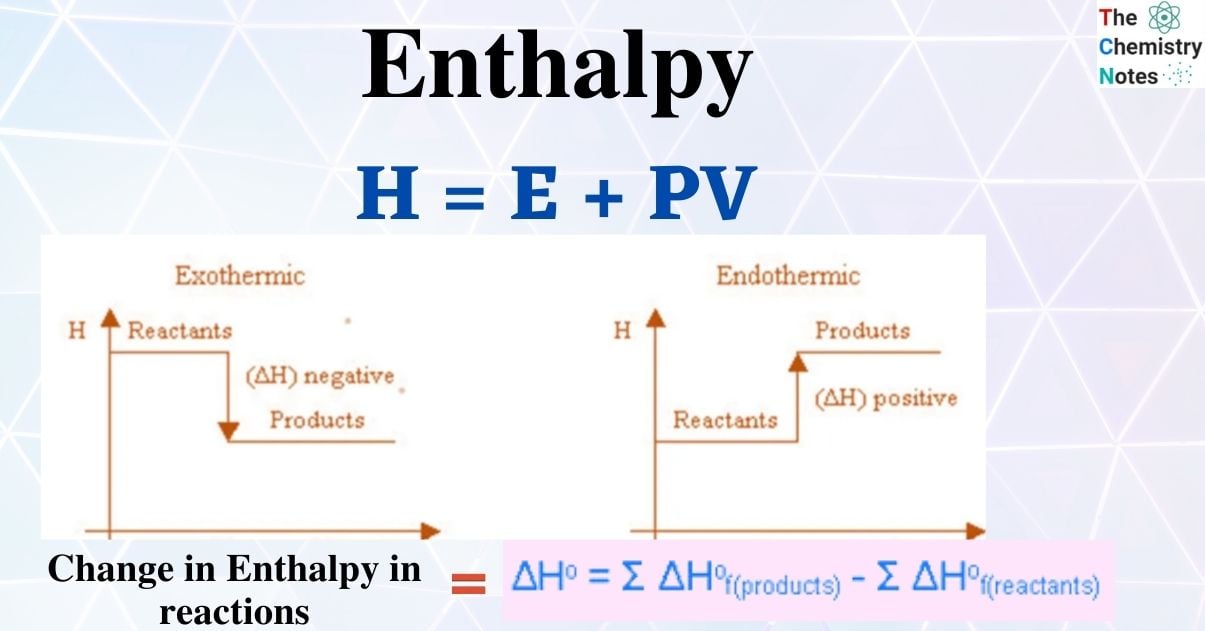
Enthalpy is one of those thermodynamic quantities that shows up everywhere in chemistry and engineering, yet most people fumble the actual computation when they hit a non-standard problem. The textbook definition is straightforward enough — enthalpy H equals internal energy plus pressure times volume, or H = U + PV — but applying it to real data requires knowing which path to take and what assumptions are actually safe to make. I spent years working in process engineering, and one of the first things I learned was that enthalpy calculations fail quietly. You can plug numbers into the right equation and still get an answer that looks plausible but is completely wrong because you used the wrong reference state or ignored a phase change that was happening at constant temperature. That lesson came hard when I was sizing a heat exchanger for a distillation column and the enthalpy balance was off by twelve percent. Turns out I had been using liquid water enthalpy values at one hundred degrees Celsius instead of steam table values, which matter enormously when you are dealing with phase transitions. The fix was switching to NIST REFPROP for all steam properties and redoing the balance, which took about forty-five minutes and revealed exactly where the error lived.
How To Calculate Enthalpy Using Calorimetry
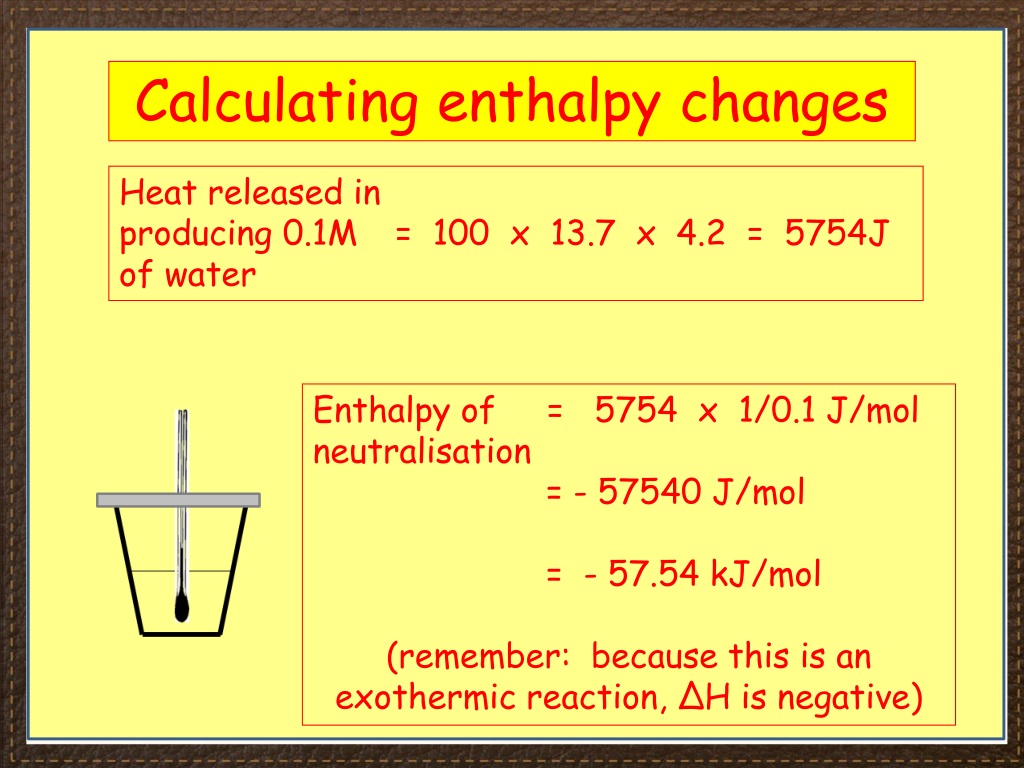
The most common way people encounter enthalpy in practice is through calorimetry. You run a reaction or physical process in an insulated container, measure the temperature change, and back-calculate the heat exchanged. The fundamental equation is q = m × c × T, where q is the heat, m is the mass of the substance absorbing or releasing heat, c is the specific heat capacity, and T is the temperature change. Here is what most guides leave out: the specific heat capacity is rarely constant across a wide temperature range. If you are working with water between twenty and eighty degrees Celsius, using a single value of four point one eight joules per gram per kelvin introduces maybe one to two percent error. But if you are working with organic solvents over a hundred degree span, the variation can push past five percent, and that matters when you are trying to balance an energy equation to within a few kilojoules. Let me walk through a concrete example. Suppose you dissolve five point zero grams of potassium chloride in one hundred grams of water at twenty-five degrees Celsius in a coffee cup calorimeter, and the temperature drops to twenty-one degrees. You want the enthalpy of solution per mole. First, calculate the heat absorbed by the solution: q = 100 g × 4.18 J/g·K × (21 - 25) K = -1672 J. The negative sign means the solution absorbed heat from the water, which makes sense because the temperature dropped. The moles of KCl is 5.0 g divided by 74.55 g/mol = 0.0671 mol. The enthalpy of solution is -1672 J / 0.0671 mol = -24.9 kJ/mol, or positive twenty-four point nine kilojoules per mole if you report it as the system absorbing heat.
Wait, I need to flag something here. That calculation assumes the calorimeter itself absorbed negligible heat. In reality, the thermometer, stirrer, and cup all have heat capacity. A proper lab setup accounts for the calorimeter constant, usually determined separately by running a known reaction like the neutralization of HCl with NaOH. Without that correction, your result could be off by ten to twenty percent depending on your apparatus. I learned this the hard way during an undergraduate lab where my calculated enthalpy of neutralization was thirty-two kilojoules per mole instead of the literature value near fifty-seven. The missing piece was the calorimeter constant of about eighty-five joules per kelvin, which I should have measured before touching the actual experiment.
Get the Full Details

Using Standard Enthalpies of Formation
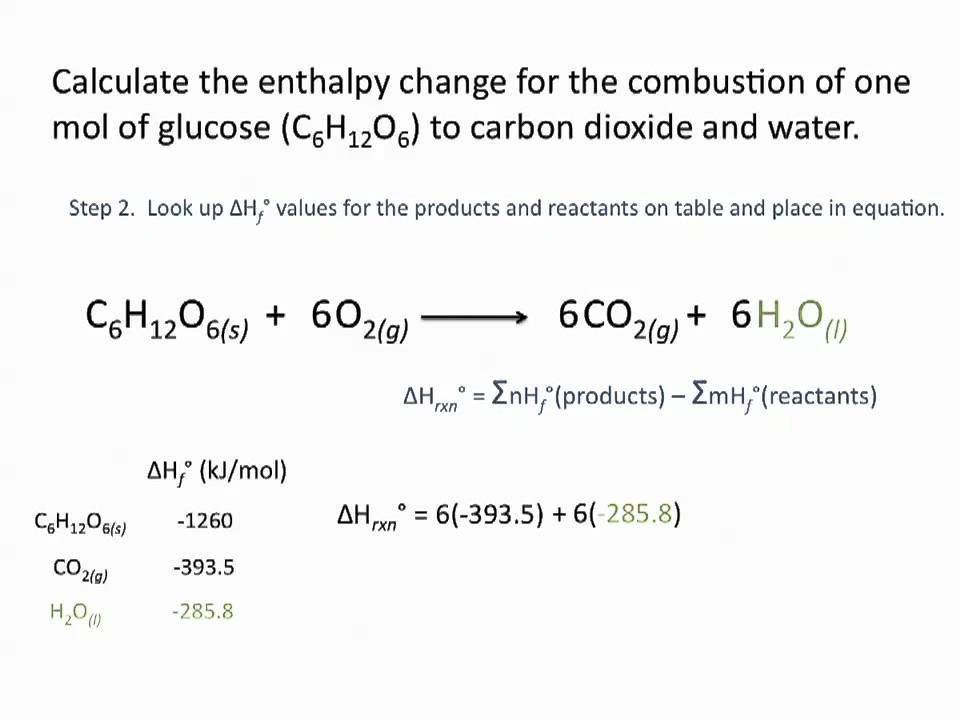
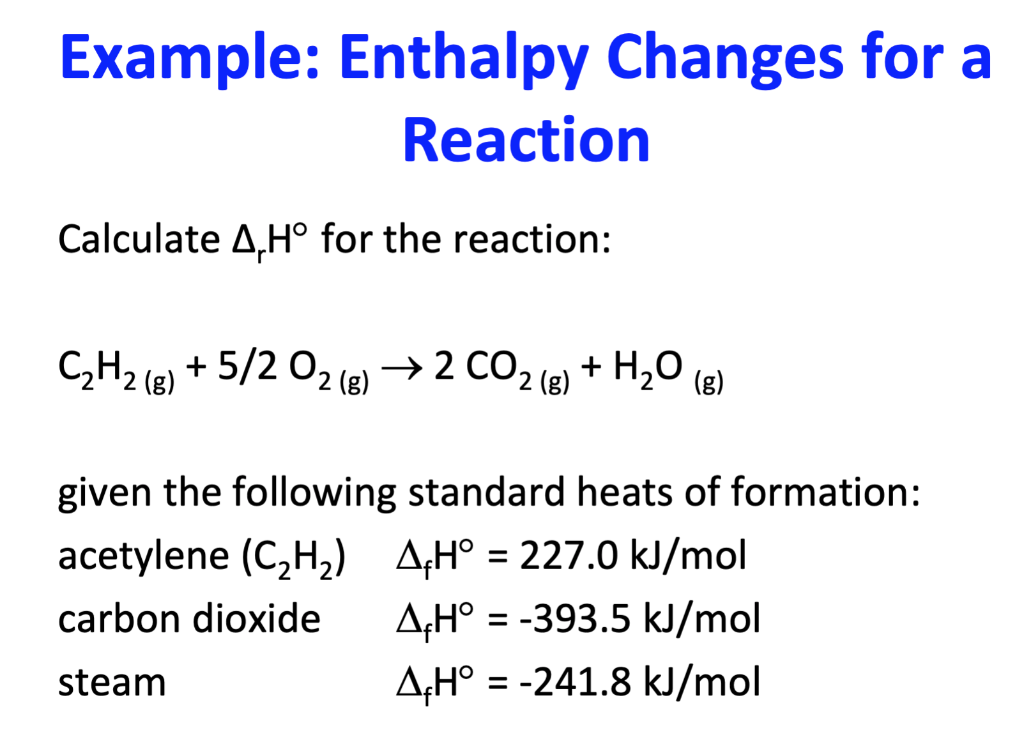
When you cannot run a calorimeter experiment, or when you need enthalpy changes for reactions that are too dangerous or impractical to measure directly, you fall back on standard enthalpies of formation. The principle is simple: the enthalpy change of a reaction equals the sum of the standard enthalpies of formation of the products minus the sum for the reactants, each multiplied by their stoichiometric coefficients. Delta H reaction = sum(n × Delta Hf products) - sum(m × Delta Hf reactants) The catch is that these values are only valid at standard conditions: one bar pressure and usually twenty-five degrees Celsius. If you are working at elevated temperatures, you need to apply a heat capacity correction using Kirchhoff's equation. The integrated form accounts for the temperature dependence of enthalpy when your reaction runs at three hundred Kelvin instead of two hundred ninety-eight.
Delta H(T2) = Delta H(T1) + integral from T1 to T2 of Delta Cp dT In practice, people approximate this by assuming Delta Cp is constant over the temperature range, which works reasonably well for small ranges but accumulates error at larger spans. I once saw a student use this approximation across a two hundred degree range and end up with an enthalpy value that was seven percent off. The fix was breaking the integration into smaller temperature increments and using polynomial fits for Cp from standard tables, which added about twenty minutes of work but brought the result into agreement with published data.
Hess's Law and Reaction Path Independence
Hess's law states that the total enthalpy change for a reaction is independent of the pathway. This is a direct consequence of enthalpy being a state function. What this means practically is that you can construct a target reaction from a series of measured reactions, add their enthalpy changes together, and get the answer for the reaction you actually care about even if you never ran it directly. This is especially valuable for reactions that are difficult to isolate. The formation enthalpy of carbon monoxide is a classic case. You cannot cleanly burn carbon to CO without also producing some CO2, so you measure the combustion of C to CO2 and the combustion of CO to CO2 separately, then subtract. The algebra works out because you are exploiting the state function property: the enthalpy difference between graphite plus oxygen and carbon monoxide is the same whether you go through carbon dioxide or not. One thing that trips people up is the sign convention when you reverse a reaction. If you flip a reaction, you must flip the sign of its enthalpy change. I have seen this mistake in at least three different textbook solutions manuals, and it propagates through entire homework sets. The rule is straightforward but easy to miss when you are juggling multiple reactions: reversing a reaction reverses the direction of heat flow relative to the system.

Bond Enthalpy Estimates
When you need a quick estimate and high precision is not required, bond enthalpies provide a useful shortcut. The idea is that breaking bonds costs energy and forming bonds releases energy, so the net enthalpy change is the sum of bond energies for bonds broken minus the sum for bonds formed. Delta H sum(bond energies broken) - sum(bond energies formed) Bond enthalpies are average values derived from many different molecules, which is both their strength and their weakness. They work well for gas-phase reactions where the bonding environment is relatively consistent, but they become unreliable when you have significant resonance stabilization, strain effects, or when the reaction involves condensed phases. The error margin is typically ten to twenty kilojoules per mole for reasonable estimates, which is fine for a quick check but insufficient for design work.
I used bond enthalpies extensively during early career work to screen reaction pathways before committing to detailed calculations. The approach let me evaluate dozens of options in a few hours instead of weeks. When I later ran proper quantum chemical calculations, the bond enthalpy estimates were usually within fifteen percent of the more rigorous results, which validated the shortcut for preliminary work. The exceptions were cases with conjugated systems or aromatic stabilization, where the error jumped to thirty or forty percent because the average bond energies do not capture delocalization effects.
Common Pitfalls and When Enthalpy Calculations Fail
The biggest source of error I see is ignoring the reference state. Enthalpy values are only meaningful relative to a defined zero point. Standard enthalpies of formation set elements in their standard states at twenty-five degrees Celsius and one bar to zero. If you mix data from different sources with different reference states, your calculation is garbage regardless of how careful you are with the arithmetic. Another frequent mistake is treating enthalpy as if it is conserved. It is not. Energy is conserved, but enthalpy can change through work interactions and heat transfer. In open systems with flowing streams, you need to account for flow work, which is why the enthalpy term appears naturally in steady-flow energy balances. I spent an afternoon debugging a mass and energy balance for a compression process where the discrepancy traced back to forgetting that the inlet and outlet were at different pressures, and the PV term was contributing several kilojoules per mole that I had neglected. Phase changes deserve special attention. The enthalpy of vaporization or fusion can be an order of magnitude larger than sensible heat effects over the same temperature interval. Water at one hundred degrees Celsius has an enthalpy of vaporization of about forty point six six kilojoules per mole, which dwarfs the sensible heat needed to raise liquid water from twenty to one hundred degrees, roughly three kilojoules per mole. Miss the phase change and your energy balance will be wildly off.
For highly non-ideal systems, particularly near critical points or with strong intermolecular interactions, simple enthalpy correlations break down. Real gases deviate from ideal behavior, and equations of state like Peng-Robinson or Soave-Redlich-Kwong become necessary. I encountered this when working with refrigerant blends at high pressures, where the ideal gas assumption introduced errors exceeding twenty percent in the enthalpy calculation. Switching to a cubic equation of state with appropriate mixing rules reduced the error to below three percent, which was the difference between a design that worked and one that did not.
Practical Computation Workflow
Here is a practical sequence I follow when tackling an enthalpy calculation: First, identify the process type: constant pressure, constant volume, flowing stream, or batch reaction. This determines which formulation is appropriate. Second, gather all relevant thermodynamic data from consistent sources. NIST Chemistry WebBook, DEAP, or appropriate handbooks are reliable starting points. Third, check for phase changes across the temperature and pressure range of interest. Fourth, apply the appropriate equation and track units carefully. Fifth, validate against a known case or limiting behavior when possible. The validation step is where most people skip and later wonder why their numbers do not match literature values. Run your calculation on a system with a known answer first. If you are computing the enthalpy of combustion for methane, compare against the published value near negative eight hundred ninety kilojoules per mole before trusting your method on an unfamiliar reaction. This usually takes less than five minutes and catches methodological errors early.
For complex industrial processes, spreadsheet-based enthalpy balances are workable for simple cases, but dedicated process simulation software like Aspen Plus or CHEMCAD becomes necessary when you have dozens of components, multiple phases, and recycle streams. I transitioned from hand calculations to Aspen for a solvent recovery system where the enthalpy balance involved fourteen organic compounds across three phases. The manual approach would have required weeks of tedious iteration, while the simulator produced a converged solution in about an hour after initial setup.
When to Use Alternative Approaches
Sometimes enthalpy cannot be measured or estimated reliably through standard methods. For exotic materials, extreme conditions, or reactions with unstable intermediates, experimental calorimetry with specialized equipment may be the only path. High-temperature drop calorimetry, solution calorimetry, and differential scanning calorimetry each have their domains of applicability. If you are working at temperatures above five hundred Kelvin, remember that heat capacity data becomes harder to find and more temperature-dependent. Polynomial fits from standard references typically extend to one thousand or twelve hundred Kelvin, but extrapolation beyond that range is unreliable. I had to estimate enthalpies for a high-temperature process running near eighteen hundred Kelvin by combining Shomate equation parameters with group contribution methods, accepting an uncertainty of maybe fifteen percent in the final result. Biochemical systems introduce additional complexity because standard states for protons differ between conventional thermodynamics and biochemistry. The biochemical standard state sets pH to seven, which shifts apparent enthalpy values compared to the chemical standard state. If you are working at the interface of chemistry and biology, make sure you are using the correct convention, or your enthalpy of reaction will be systematically wrong by an amount that depends on the proton stoichiometry of your process.
The bottom line is that enthalpy calculation is straightforward when the problem is simple and the data is clean. Complications arise from non-ideal behavior, phase transitions, temperature-dependent heat capacities, and inconsistent reference states. Track those carefully, validate against known cases, and you will avoid most of the errors that trip up people who treat enthalpy as just another number to plug into an equation.