The Basics of Partial Pressure
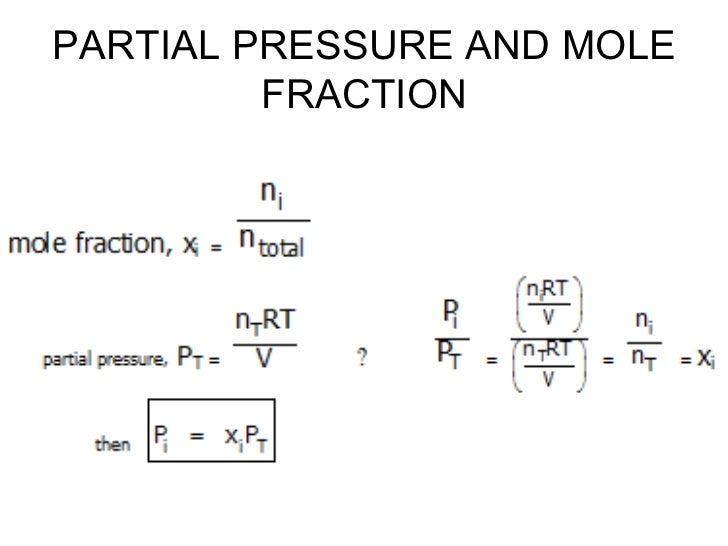
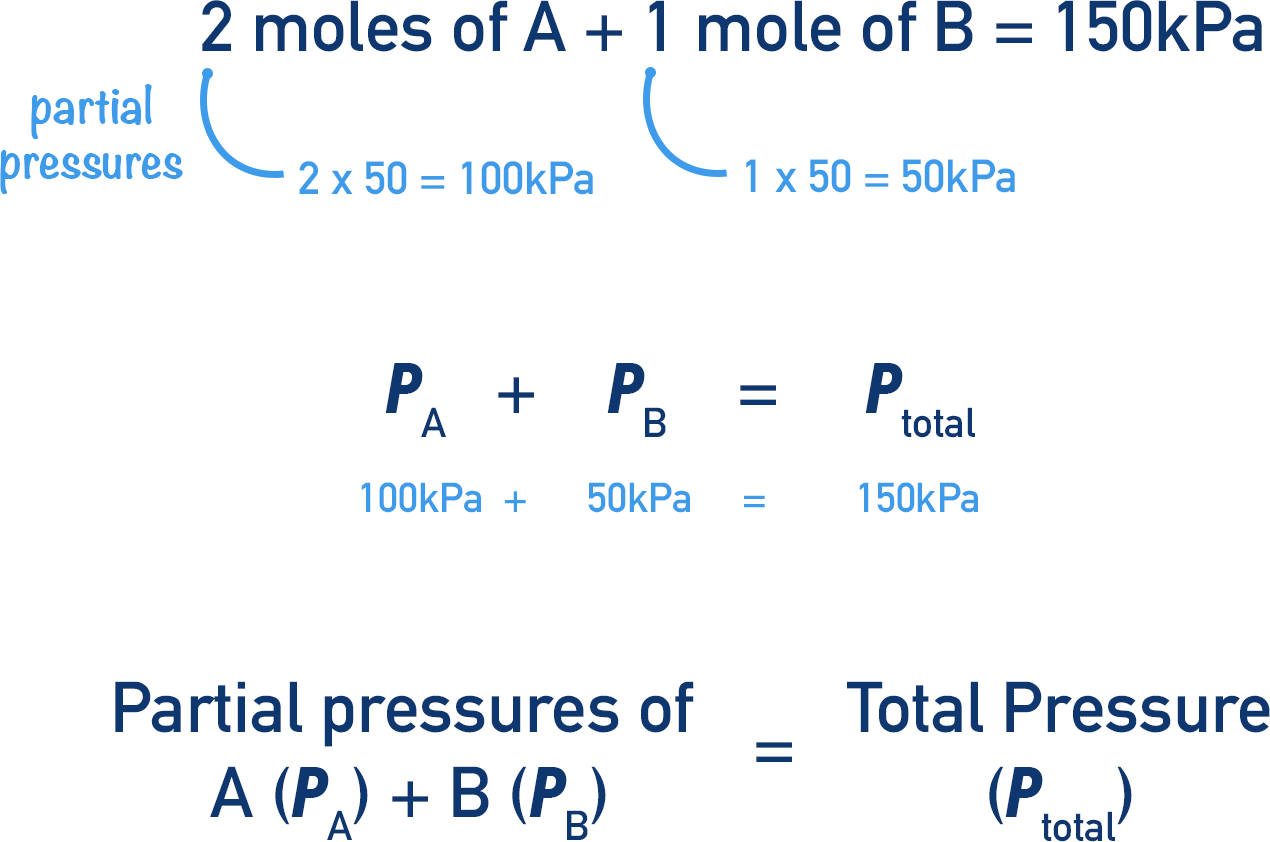
Partial pressure is just the pressure that one gas in a mixture would exert if it were alone in the same volume at the same temperature. That's it. It's not some mystical concept—it's Dalton's Law, and it's been around since 1801. The formula is straightforward: P_total = P_1 + P_2 + P_3 + ... + P_n. Each individual gas contributes its own pressure to the total, and the sum of those contributions equals whatever the total pressure reads on your gauge. The way you actually get there in practice is through mole fraction. You take the number of moles of your target gas, divide it by the total moles of every gas in the mixture, and multiply that ratio by the total pressure. So P_gas = (n_gas / n_total) × P_total. The mole fraction is dimensionless, which means your units for partial pressure end up being whatever pressure units your total pressure was already in—kPa, atm, bar, mmHg, doesn't matter as long as they're consistent. Here's where people mess this up immediately: they confuse mole fraction with volume fraction or mass fraction. In an ideal gas mixture, mole fraction and volume fraction are actually the same thing, but mass fraction is completely different and using it by accident will throw your answer off by however much the molecular weights differ. I've seen it happen in lab reports more than once. Nitrogen and oxygen have close enough molar masses that the error looks small, but pair something like hydrogen with carbon dioxide and the mistake is glaring.
How To Calculate Partial Pressure in Real Conditions
The standard approach assumes ideal gas behavior, which works fine for most atmospheric and laboratory scenarios at moderate pressures. But when you're working above 10 bar or dealing with gases that have strong intermolecular forces, the ideal approximation starts to drift. In those cases you need fugacity coefficients, which adjust the partial pressure to account for non-ideality. For a quick check, the compressibility factor Z tells you whether you're in the ballpark—anything below 0.95 or above 1.05 and you should probably be thinking about corrections. I ran into this exact problem last year while designing a pressurized gas blending station for an industrial welding setup. We were mixing argon, CO2, and helium at around 15 bar for a process gas blend. The spec called for precise partial pressures of each component because the arc characteristics depended on it. At first I just ran the ideal gas calculation and got numbers that looked right on paper. Then I ran them through the NIST REFPROP database and the CO2 partial pressure was about 4% higher than my ideal calculation predicted. Four percent sounds small until you're tuning a welding process and the bead quality is inconsistent across shifts. The workaround was to use the virial equation of state for the initial blend and then verify with a real fluid property program. For CO2 at those conditions, the second virial coefficient correction was sufficient—I didn't need the full equation of state. If you don't have access to REFPROP or similar software, the Redlich-Kwong or Peng-Robinson equations give decent estimates without the overhead. My rule of thumb is that for binary or ternary mixtures at moderate pressures, a single virial coefficient correction gets you within about one percent, which is usually good enough for anything except analytical chemistry work.
Another thing that bites people is assuming partial pressure stays constant when you change temperature. It doesn't. If you seal a gas mixture in a rigid container and heat it, the total pressure goes up according to the ideal gas law, but the mole fractions don't change, so each partial pressure scales linearly with absolute temperature. Double the Kelvin temperature and every partial pressure doubles too. This matters when you're specifying equipment ratings—a tank rated for a certain pressure at room temperature might see substantially higher partial pressures of individual components at elevated temperatures, even if the total pressure seems manageable. For water vapor specifically, partial pressure is essentially the same thing as vapor pressure when you have a saturated mixture. That's why humidity meters often report in millibars of water vapor partial pressure rather than relative humidity—the absolute number is more useful for process calculations. The Antoine equation or the Tetens formula can give you the saturation vapor pressure at a given temperature, and from there you can find the actual partial pressure by multiplying by the relative humidity as a decimal. A few practical notes about the common errors: Make sure all pressures are in the same units before adding them. I can't stress this enough—mixing mmHg with kPa and not converting first is the most frequent mistake in any setting where people calculate partial pressures casually. Second, remember that partial pressure only applies to the gaseous phase. If a component condenses out at your conditions, its partial pressure caps at the saturation vapor pressure and won't increase further no matter how much more of that substance you add. This is especially relevant for water in compressed air systems and for organic solvents in any kind of headspace analysis.
Get the Full Details
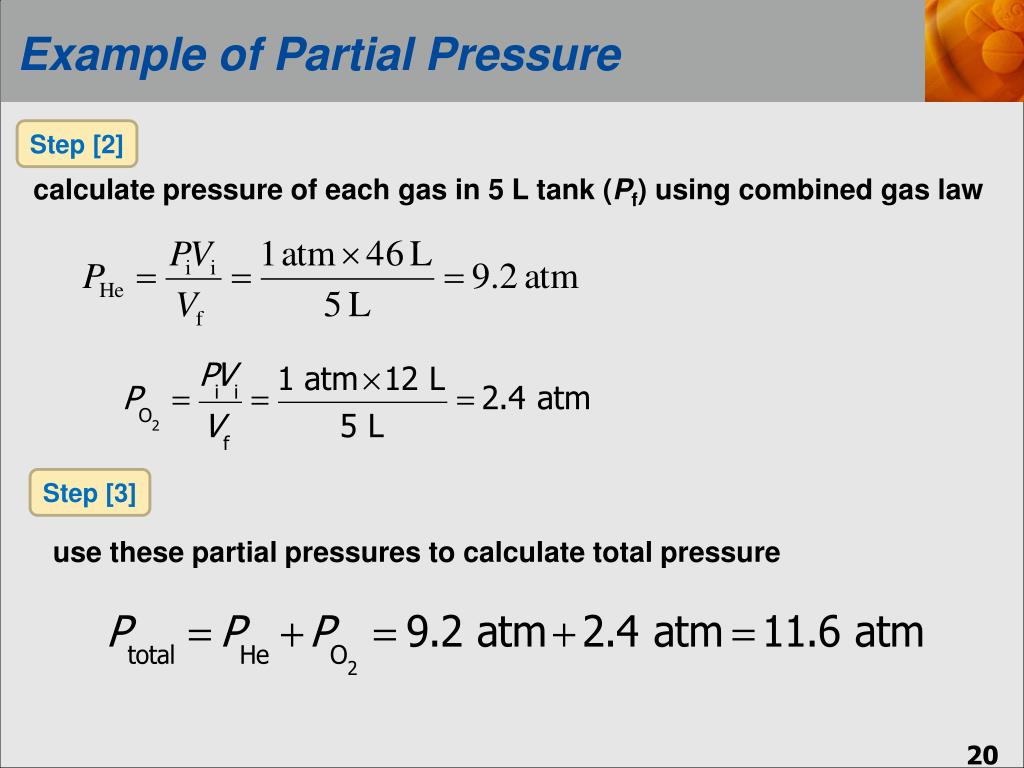
When you're dealing with gas permeation through membranes or diffusion through tissues, the driving force is the partial pressure gradient, not the concentration gradient. They're related through Henry's law, but in practice engineers and physiologists work directly in pressure units because it's cleaner. If you find yourself converting back and forth between concentration and partial pressure constantly, you're probably overcomplicating something. The shortcut of using percentage by volume directly as the mole fraction only works cleanly for ideal gases. Air at sea level? Fine. A high-pressure reactor with supercritical CO2? Not fine. Know your regime before you apply the shortcut.