The Basics Nobody Tells You
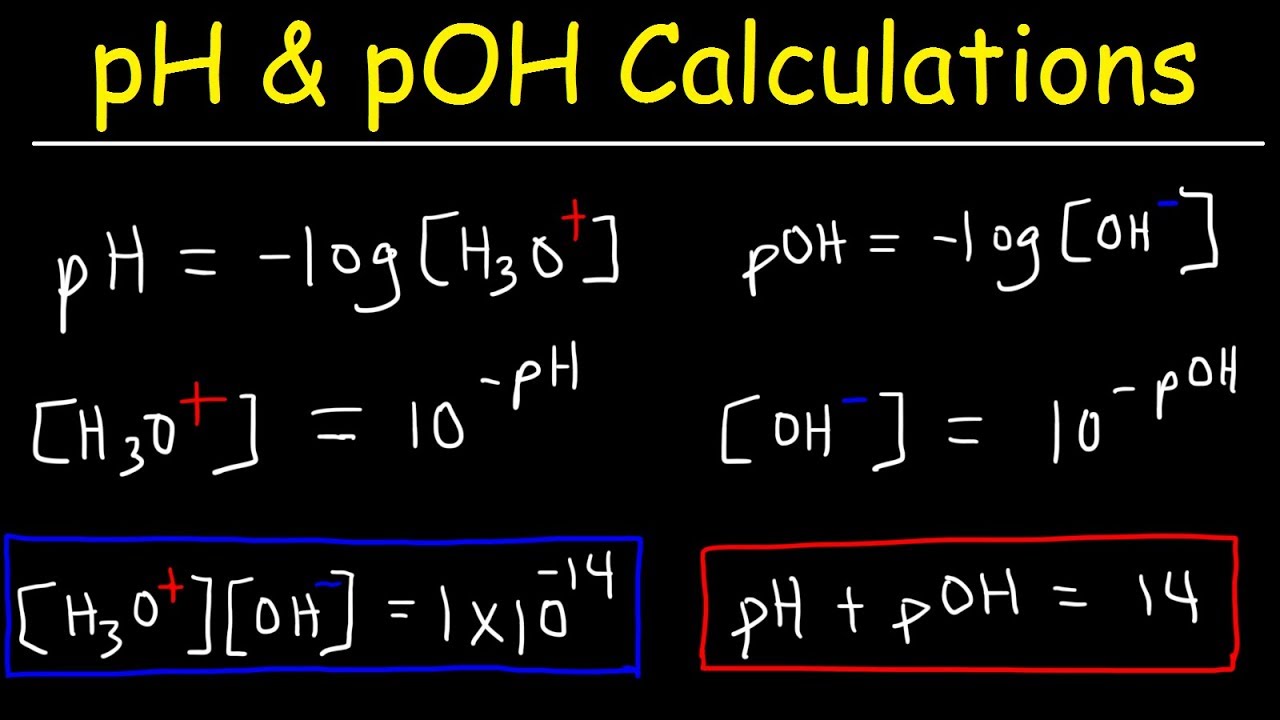
pH is just negative log base 10 of the hydrogen ion activity. The textbook says pH equals -log[H+], but in practice the difference between concentration and activity matters a lot once you step away from dilute solutions. I learned that the hard way when I was working with a wastewater sample that had an ionic strength well over 0.1 molal. My initial readings using the simple concentration formula were giving me consistent offsets of about 0.2 to 0.4 pH units compared to what the calibrated bench meter showed. Once I switched to calculating activity coefficients using the Davies equation, everything clicked into place. The core formula is still straightforward: pH = -log10(aH+), where aH+ is the activity of the hydrogen ion. Activity equals concentration multiplied by the activity coefficient gamma. So pH = -log10(gamma × [H+]). For most lab work at low ionic strength, gamma is close to 1 and you can basically ignore it. But that approximation falls apart fast.
How To Compute For Ph in Real Conditions
Here is the practical workflow I use when I need to calculate pH from scratch instead of just reading it off a meter. First, determine the concentration of all ions in solution. This sounds obvious but people skip it. If you are dealing with a strong acid like HCl at 0.01 M, the hydrogen ion concentration is essentially 0.01 M because strong acids dissociate completely. For weak acids it is different and you need an equilibrium calculation. For a weak acid such as acetic acid at 0.1 M, you set up the Ka expression. Acetic acid has a Ka of about 1.8 × 10^-5. You solve x² / (0.1 - x) = Ka for x, where x is [H+]. Since Ka is small, x turns out to be roughly 1.34 × 10^-3 M, giving a pH around 2.87. Do not use the simplified version that assumes 0.1 minus x is basically 0.1 unless you actually check that the approximation holds. In borderline cases where the acid is moderately concentrated or Ka is larger, dropping the x term introduces a meaningful error. Next comes the activity coefficient. I usually calculate ionic strength first using I = 0.5 × sum(ci × zi²) for all ions present. Then I apply the Davies equation: log(gamma) = -0.509 × z² × [sqrt(I) / (1 + sqrt(I)) - 0.3 × I]. For monovalent ions at moderate ionic strength this gives gamma values in the 0.7 to 0.9 range. Multiply your hydrogen ion concentration by gamma to get activity, then take the negative log. That is the actual pH you should expect in that solution.
Strong bases follow the same logic in reverse. Find [OH-], convert to pOH, then subtract from 14 at 25 degrees Celsius. But remember that 14 is temperature dependent. At 37 degrees Celsius, neutral pH is closer to 6.8, not 7. I have seen people calculate pH for blood or physiological samples using the 25-degree assumption and then wonder why their numbers looked off compared to clinical reference ranges. The temperature correction is not optional when you care about accuracy. For polyprotic acids like phosphoric acid, you need to handle multiple dissociation steps. Phosphoric acid has three Ka values: 7.5 × 10^-3, 6.2 × 10^-8, and 4.2 × 10^-13. In a 0.1 M solution, the first dissociation dominates and the second contributes only a small additional amount of H+. You can usually approximate it by solving the first equilibrium, then using that result as the starting point for the second. The contribution from the third dissociation is negligible for almost any practical purpose. I used to set up full simultaneous equations for these, which took forever and gave no better answer than the sequential approximation. A colleague pointed out that the Debye-Hückel based activity corrections were doing more damage to the accuracy than my messy math, so now I just stick to the simpler sequential approach and focus my effort on getting the ionic strength right. One edge case that caught me last year was calculating pH for a solution containing both a weak acid and its conjugate base at similar concentrations. The Henderson-Hasselbalch equation seems perfect here: pH = pKa + log([A-]/[HA]). But I was working with a phosphate buffer in a high-salt matrix and the measured pH kept drifting after I prepared it. The problem was not the equation itself but the activity coefficients changing as salts precipitated and re-dissolved over time. I ended up having to measure the ionic strength repeatedly and recalculate gamma rather than assuming it stayed constant. In the end I just calibrated the meter directly against standard buffers prepared in the same high-salt medium, which was faster and more reliable than trying to compute it from first principles every time.
Get the Full Details
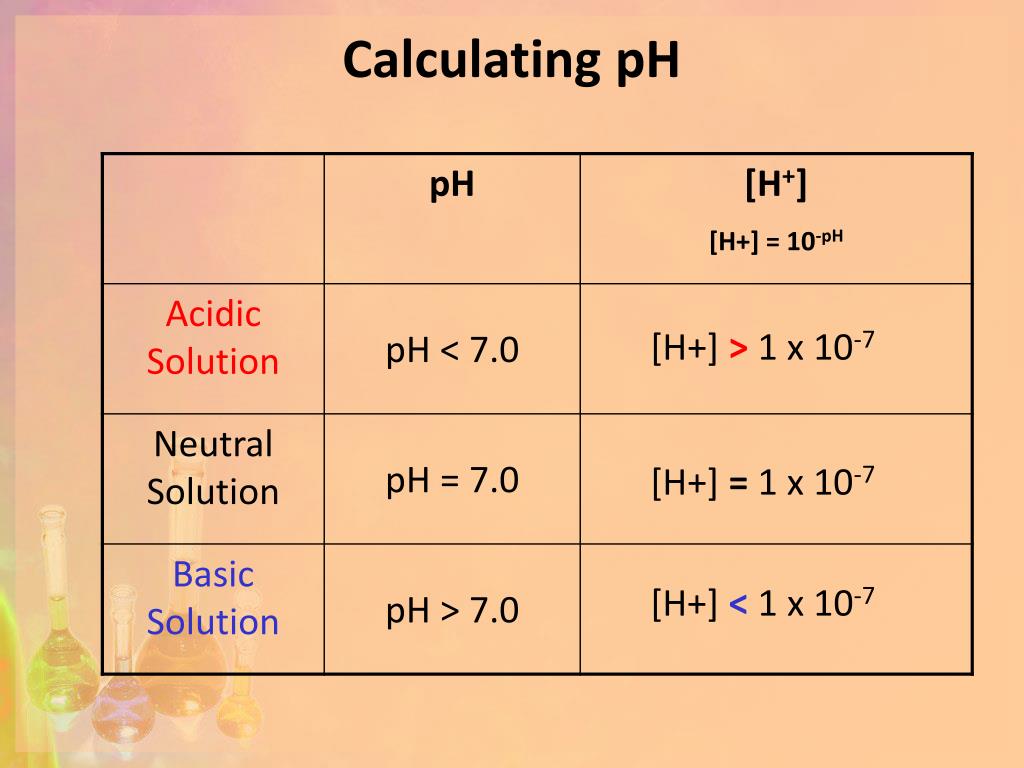
There are limits to computation-based pH estimation that you should accept upfront. The method breaks down in solutions with very high ionic strength above about 0.5 M, where even the Davies equation becomes unreliable. It does not account for ion pairing or complex formation, which matters in environmental and biological samples containing metals and organic ligands. Temperature effects beyond a simple Kw shift are difficult to model without empirical data. And for mixtures of multiple weak acids and bases, the algebra gets unwieldy quickly. When computation is not enough, iterative numerical methods or specialized software like PHREEQC or Visual MINTEQ handle the equilibrium calculations properly. They model speciation, activity corrections, and precipitation simultaneously. The learning curve is steeper, but they save hours of manual work on complex systems. For routine laboratory work with simple acid-base solutions, the manual method I described above is perfectly adequate and takes roughly five to ten minutes per sample once you know the steps. The time you save by understanding the underlying chemistry shows up when your calculated values stop matching your meter readings and you actually know which assumption to adjust.