The Real Way Enthalpy Works
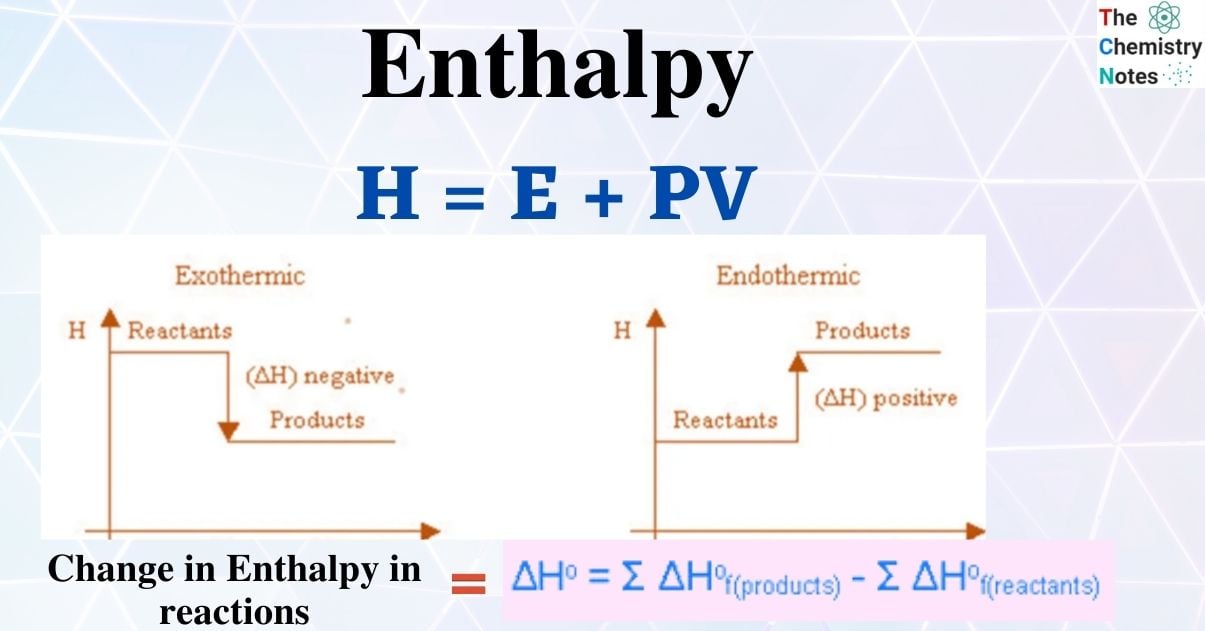
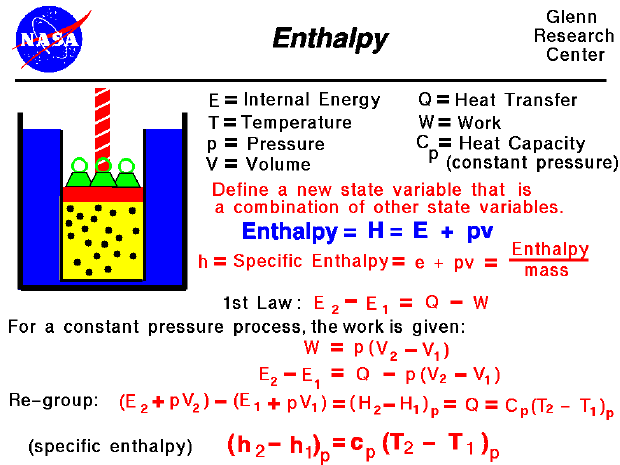
Most people encounter enthalpy in a thermodynamics class and immediately get bogged down in abstract definitions. The equation H = U + PV looks clean on paper but doesn't tell you much about when or why to actually use it. I spent years watching students trip over this same material, and honestly, the problem is almost always a mismatch between the formula and the physical situation they're trying to solve. Enthalpy is the total heat content of a system at constant pressure. That's the textbook answer. In practice, it's a bookkeeping tool that lets you track energy changes without worrying about the work the system does on its surroundings. When you're dealing with open systems like reactors or heat exchangers running at steady pressure, enthalpy is far more useful than internal energy. Period.
How To Figure Out Enthalpy in Real Problems
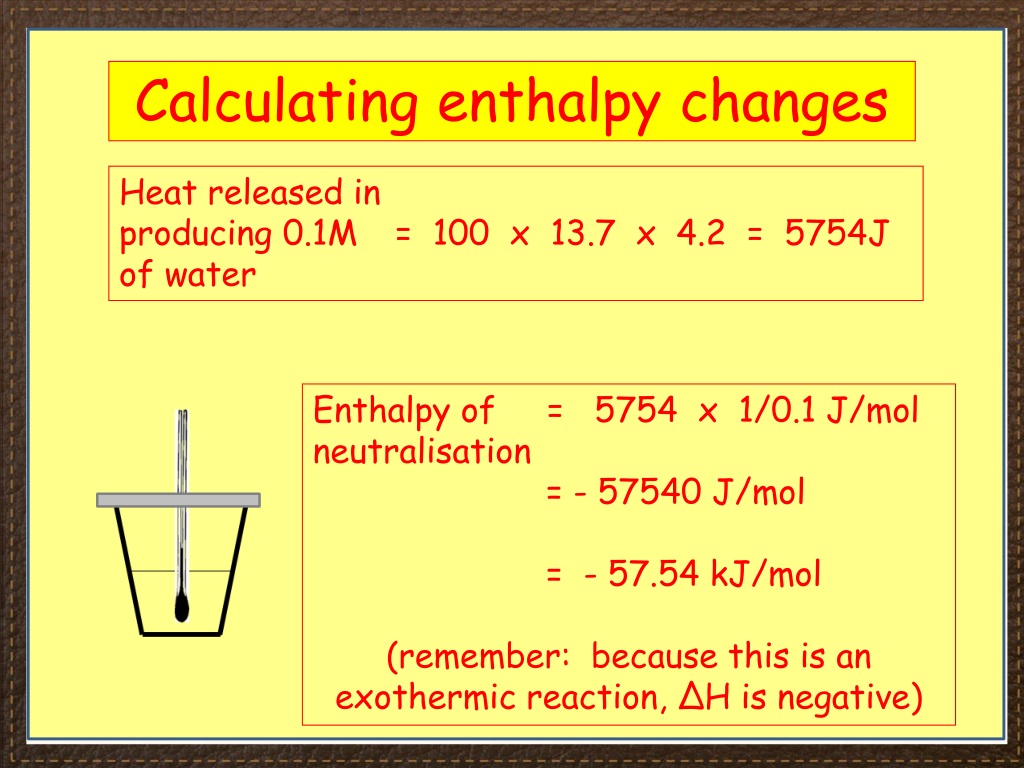
Start by identifying whether the process happens at constant pressure or constant volume. This single decision determines your entire approach. If pressure is constant, you work directly with enthalpy. If volume is constant, you stick to internal energy and convert later if needed. I can't count how many times I've seen someone plug numbers into a heat capacity equation when they should have been tracking phase changes instead. The method changes completely depending on what's actually happening in the system. For sensible heat calculations, the basic formula is straightforward. Q equals mass times specific heat capacity times the temperature change. Use Cp for constant pressure and Cv for constant volume. The difference between them matters mostly for gases. For liquids and solids, Cp and Cv are close enough that you can usually treat them as identical without introducing significant error into your calculations. When phase changes are involved, stop using temperature differences entirely. Enthalpy of fusion and enthalpy of vaporization are separate terms you add to your calculation. During melting or boiling, temperature stays constant while energy continues to flow into the system. If you try to use Q equals mCpDeltaT across a phase boundary, you will get the wrong answer. I learned this the hard way during my first year working on a distillation column simulation where I accidentally calculated a reboiler duty that was off by nearly forty percent because I had missed a latent heat term in the mixture. Took me three hours to trace the error back to that single omission.
For gas phase processes where pressure and temperature both change, you need to be more careful. The full expression involves integrating Cp over the temperature range and then adding a pressure correction term if the gas deviates from ideal behavior. Most undergraduate problems assume ideal gas behavior, which simplifies things considerably. Under those conditions, enthalpy depends only on temperature for an ideal gas. Pressure changes don't affect it. That is one of those counter-intuitive facts that trips people up constantly. A gas expanding through a valve at constant enthalpy, the Joule-Thomson effect, will change temperature precisely because real gases deviate from this assumption. If your problem involves high pressures or low temperatures where ideal gas assumptions break down, you need an equation of state or tabulated data rather than a simple Cp integral.
Get the Full Details

Common Pitfalls That Wreck Calculations
Units are the most common source of errors. Cp values vary wildly depending on whether they are given per mole or per gram, and whether the temperature unit is Celsius or Kelvin. A Cp of 4.18 J per gram Kelvin for water is fine until you multiply it by a molar mass and realize you need the value per mole instead. Always verify that your mass, your heat capacity, and your temperature change are expressed in consistent units before you start multiplying numbers together. This mistake alone accounts for roughly half the wrong answers I see in homework submissions. Another issue people overlook is reference states. Enthalpy values are relative, not absolute. Tables list enthalpy relative to some arbitrary reference point, usually the liquid phase at a standard temperature and pressure. When you calculate an enthalpy change between two states, the reference state cancels out. But if you are combining enthalpy values from different sources or different tables, the reference states might not match. I ran into this exact problem when cross-referencing steam table data with a chemical engineering handbook for a boiler efficiency calculation. The two sources used different reference points for liquid water, and my final answer was shifted by exactly the enthalpy of vaporization at the reference temperature. Once I aligned both datasets to the same baseline, the discrepancy disappeared immediately. For mixtures and solutions, the situation gets messier. You cannot simply add the enthalpies of pure components and expect the right answer. There are mixing enthalpies, sometimes called heats of solution, that need to be accounted for separately. In dilute aqueous solutions these effects are small, but in concentrated acid mixtures or organic solvent blends they can dominate the energy balance. If your problem involves mixing anything other than ideal gases, look for experimental data or use an activity coefficient model rather than assuming zero enthalpy of mixing.
When the Standard Approach Fails
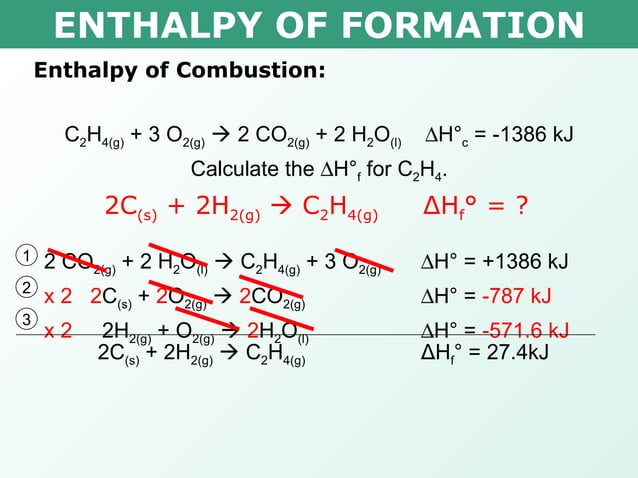
Constant pressure enthalpy calculations work beautifully for closed systems undergoing simple heating or cooling with no phase changes. They also work well for steady-flow open systems like heat exchangers, turbines, and compressors, where the steady flow energy equation reduces to an enthalpy balance. But there are situations where this framework starts to break down. Adiabatic flames are one example. The temperature you calculate from a simple enthalpy balance can exceed realistic limits if you do not account for dissociation at high temperatures. At flame temperatures above about two thousand Kelvin, molecules start breaking apart, and that endothermic process absorbs energy that a basic calculation would otherwise assign to sensible heat. If you are modeling combustion, you need equilibrium composition data or a dedicated chemical kinetics package, not just Cp values and an enthalpy balance. Another case where enthalpy alone is insufficient is when entropy generation matters. Enthalpy tells you about energy conservation, but it says nothing about whether a process is feasible or irreversible. A heat exchanger design based solely on enthalpy balances might suggest a configuration that violates the second law. I once reviewed a proposal for a waste heat recovery system that looked perfectly reasonable on an enthalpy basis but would have required heat to flow from a colder stream to a hotter one without any external work input. Enthalpy balances allowed it. Entropy analysis revealed the impossibility within minutes. Always run an entropy check alongside your enthalpy work if you are designing real equipment. For reactive systems, you need standard heats of formation in addition to sensible heat and phase change terms. The enthalpy of reaction at standard conditions is calculated from the difference between the enthalpies of formation of products and reactants. Then you adjust for the actual operating temperature using heat capacity integrals. This is the standard procedure, but it assumes complete reaction. If your conversion is less than one hundred percent, you need to account for the unreacted species carrying enthalpy out of the system as well. The energy balance is not just about what reacted. It is about everything that entered and everything that left.
Practical Workflow for Typical Problems
Here is the sequence I go through when I encounter a new enthalpy problem. First, draw a boundary around the system and label every inlet and outlet stream with their known properties. Temperature, pressure, composition, and flow rate. Second, identify what is happening in each stream. Is there a phase change? A chemical reaction? Both? Third, choose your reference state and stick with it throughout the entire calculation. Fourth, write the energy balance. For steady flow systems this is usually enthalpy in equals enthalpy out plus heat transferred plus work done. Fifth, look up or calculate the enthalpy of each stream relative to your reference state. Sixth, solve for the unknown. If you have more unknowns than equations, you are missing a constraint. Go back and check your assumptions. Heat capacity data is typically available as a polynomial function of temperature in the form of Cp equals A plus B times T plus C times T squared plus D divided by T squared. These coefficients are tabulated in standard references for hundreds of common substances. Memorizing them is unnecessary. Knowing where to find them and how to integrate the expression is what matters. The integration is mechanical. You just need to be careful with the units on the temperature variable, which is usually expressed in Kelvin in these correlations. For quick estimates, especially in the early stages of a design where you do not need high precision, constant average heat capacities over the temperature range of interest are often sufficient. Pick a mean temperature, look up Cp at that point, and proceed. This approximation introduces errors on the order of a few percent for most common substances over moderate temperature ranges. If you need better accuracy, integrate the polynomial expression instead. The extra effort is minimal with a spreadsheet, and the improvement in accuracy is noticeable when temperature spans exceed a few hundred degrees.

Steam tables remain the gold standard for water and steam calculations. No polynomial correlation comes close to the accuracy of the IAPWS formulations behind modern steam tables. If your problem involves water in any phase, use the tables or an IAPWS-based calculator. Do not attempt to approximate water properties with generic heat capacity equations unless you are doing a rough order of magnitude estimate and the consequences of error are negligible. The enthalpy of steam at typical power plant conditions can differ from a simple Cp-based calculation by several percent, and in the two-phase region the concept of Cp breaks down entirely anyway. The bottom line is that figuring out enthalpy comes down to understanding what the quantity represents, choosing the right reference state, and being systematic about accounting for every energy term in your balance. The math itself is straightforward calculus or arithmetic depending on the complexity of the problem. The difficulty is in making sure you have not overlooked a phase change, a reaction, or a mixing effect. Those are the things that sink calculations, not the equations themselves.