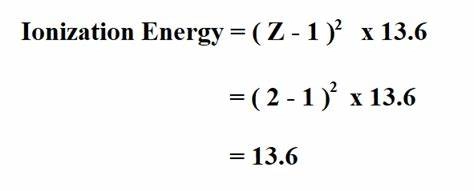
Ionization Energy Isn't as Simple as Looking It Up
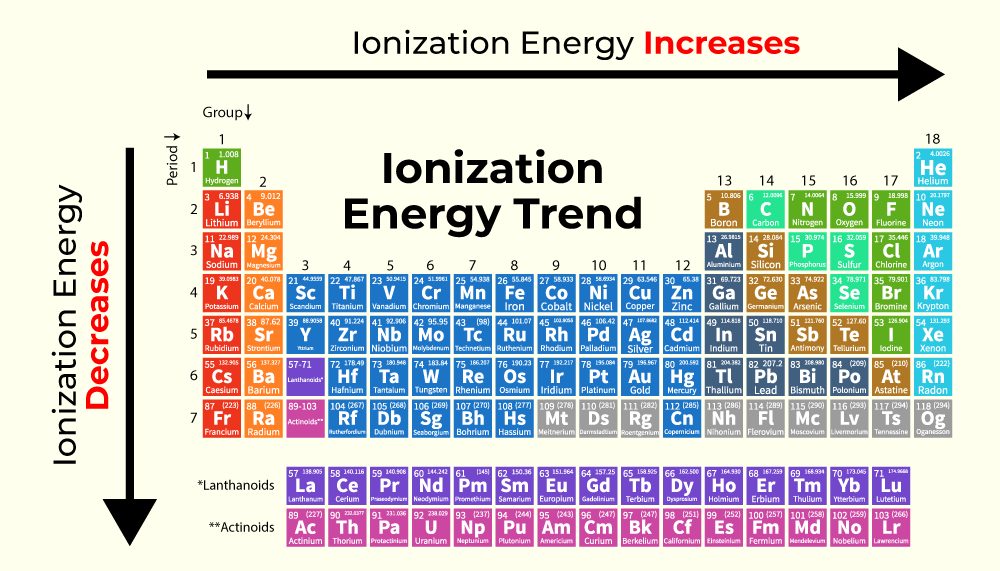
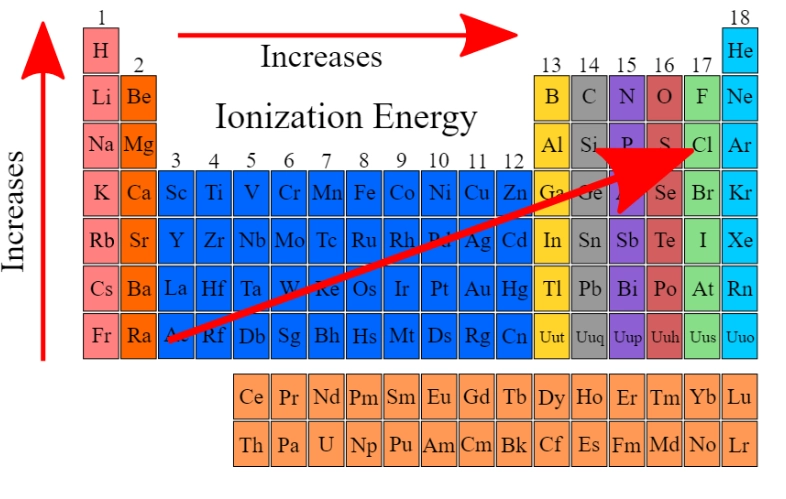
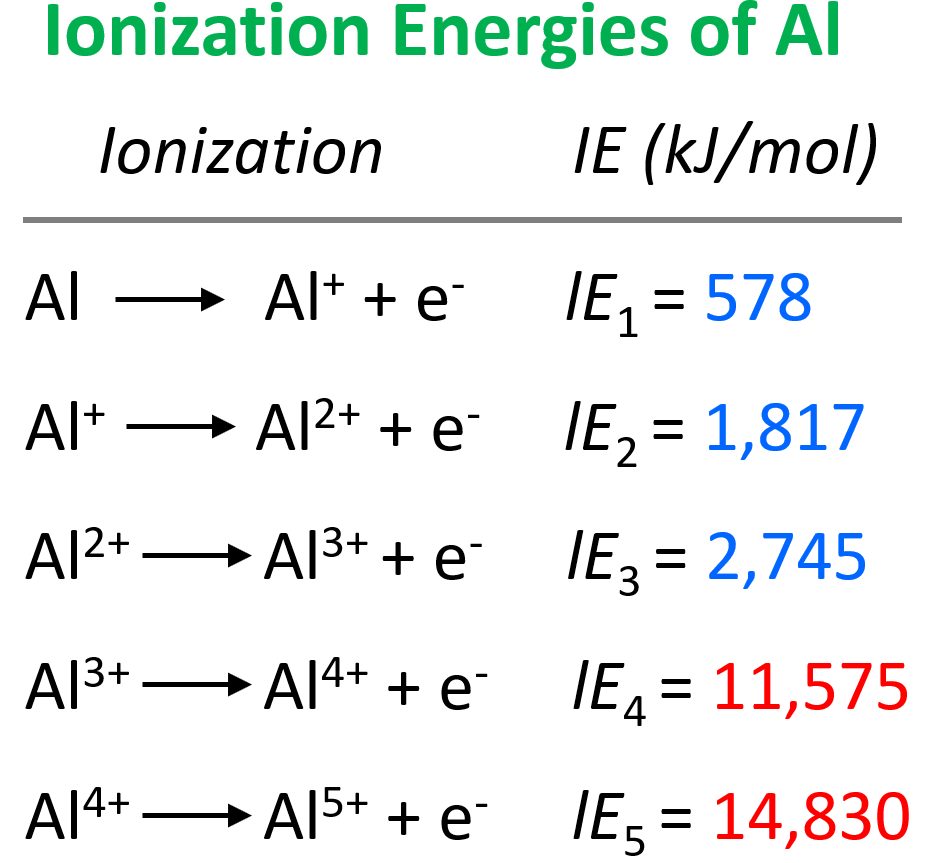
Most people learn that ionization energy increases across a period and decreases down a group, then move on without really understanding how you actually determine these values in practice. That periodic trend stuff is useful for quick estimates, but if you're working on something that requires actual numbers—whether that's computational chemistry, materials science, or just doing advanced homework correctly—you need to know the methods and their limitations. The first thing to understand is that ionization energy isn't one single thing you calculate from a single formula. It depends on which electron you're removing, what element you're dealing with, and how precise you need to be. The most straightforward definition is the minimum energy required to remove the most loosely bound electron from a neutral gaseous atom in its ground state. That's the first ionization energy. Removing a second electron requires the second ionization energy, and so on. The jump between successive ionization energies tells you a lot about electron configuration. I remember working through a problem where I needed to figure out why the third ionization energy of aluminum was roughly five times larger than the second. The standard explanation is that after removing two electrons, you've stripped away the 3p and one 3s electron, and now you're pulling from a filled 2p subshell that's much closer to the nucleus and shielded less effectively. Knowing that pattern saved me from trying to force a calculation that wouldn't work.
Experimental Methods
Photoelectron spectroscopy is the go-to technique for measuring ionization energies directly. You hit a sample with high-energy photons—usually ultraviolet or X-ray—and measure the kinetic energy of the ejected electrons. The ionization energy is the difference between the photon energy and the electron's kinetic energy. This gives you binding energies for individual electron shells, which is why it's so much more informative than just looking at a table value. The resolution depends on your photon source and detector setup, but modern instruments can resolve differences down to a few meV. For hydrogen and hydrogen-like ions, you can use the Rydberg formula: IE = R_H × Z² / n², where R_H is approximately 13.6 eV, Z is the nuclear charge, and n is the principal quantum number of the electron being removed. This works beautifully for H, He, Li², and so on. It breaks down immediately for multi-electron atoms because electron-electron repulsion and shielding complicate things in ways the simple formula doesn't account for. The Franck-Hertz experiment, while historically important, is more about confirming quantized energy levels than measuring precise ionization energies. Don't rely on it for actual numerical work.
Theoretical Calculations
For atoms beyond hydrogen, you need computational methods. Hartree-Fock theory gives you a starting point by treating each electron as moving in the average field of all the others. It's not great for correlation energy, but it's a reasonable approximation for qualitative work. Configuration interaction and density functional theory are better choices when you need numbers that actually match experimental data. DFT is widely used because it's computationally efficient, but you have to pick your functional carefully—B3LYP and PBE give different results, and the spread between them can be several eV for ionization energies. Here's something that catches people off guard: the ionization potential theorem in DFT says that the negative of the highest occupied Kohn-Sham eigenvalue equals the exact ionization energy. In practice, approximate functionals don't satisfy this exactly, and the error varies depending on the system. If you're running calculations and your ionization energy comes out 2-3 eV off from the literature value, that's often the issue, not a bug in your code.
Get the Full Details
Common Pitfalls
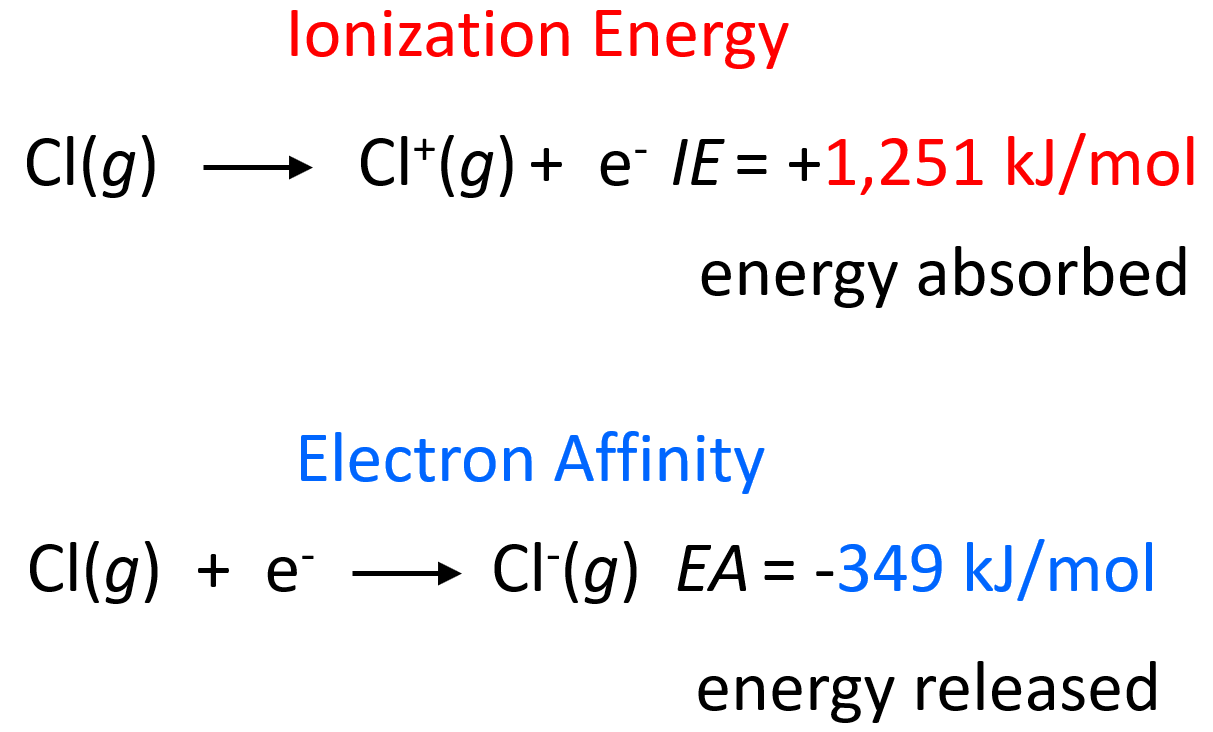
One frequent mistake is treating ionization energy and electron affinity as related quantities. They aren't. Ionization energy involves removing an electron from a neutral atom. Electron affinity involves adding an electron to a neutral atom. They measure fundamentally different processes and can have very different magnitudes for the same element. Fluorine, for example, has a higher first ionization energy than chlorine, but chlorine has a more negative electron affinity. The periodic trends don't mirror each other. Another trap is assuming ionization energy scales inversely with atomic radius in a straightforward way. It's related, but nuclear charge and shielding play equally important roles. Beryllium has a higher ionization energy than boron despite being smaller, because removing an electron from boron's 2p orbital is easier than from beryllium's fully-filled 2s subshell. The stability of filled subshells creates local exceptions to the general trend that students often miss. When you're dealing with transition metals, the picture gets even messier. The 4s and 3d orbitals are close enough in energy that the order of removal isn't always what you'd expect from the Aufbau principle. Ionization from transition metals often pulls from the 4s orbital before the 3d, which is the opposite of the filling order. This inconsistency annoys people who want clean rules, but it's just how the physics works.
When Table Values Fail You
I once had to work with a sample that wasn't pure—it contained trace impurities that fluoresced under the UV source in our photoelectron spectrometer. The background signal was masking the actual ionization peaks for the element I was interested in, and it took me about three hours to realize the problem wasn't in my calibration. Running a blank measurement and subtracting the background noise is something you should always do, but it's easy to overlook when you're rushing. If your measured ionization energies are systematically shifted, check your reference standard first. Argon's first ionization energy is well-established at 15.76 eV, and it's commonly used for calibration. If your instrument reads Ar at 15.5 eV, you've got a 0.26 eV offset to correct for. There's also the issue of phase state. Standard tabulated ionization energies refer to isolated gaseous atoms. If you're working with a solid or a solution, the environment changes everything. Solvation effects can shift ionization energies by several eV. Lattice effects in solids introduce band structure considerations that make the whole concept of a discrete ionization energy less meaningful. Don't apply gas-phase values to condensed-phase problems without adjusting for the environment, or your calculations will be wrong.
Successive Ionization Energies and What They Tell You
The pattern of successive ionization energies is actually one of the most useful tools for figuring out electron configuration from experimental data. Each successive ionization energy is larger than the previous one because you're removing electrons from an increasingly positive ion. But the real insight comes from the jumps. When the jump between two successive values is dramatically larger than the gradual increase before it, that tells you you've just broken into a new, more tightly bound shell. Sodium's first ionization energy is 496 kJ/mol. The second is 4562 kJ/mol. That's a nine-fold increase, and it's the telltale sign that the first electron came from the 3s valence shell while the second would have to come from the neon-like 2p core. This pattern is how you confirm valence electron counts experimentally. It's also why compounds like NaCl exist and NaCl doesn't—the energy cost to remove that second electron from sodium is simply too high to be compensated by bond formation.
Where the Methods Break Down
No single method covers all cases. Photoelectron spectroscopy works well for gases and some solids, but it struggles with reactive samples that degrade under UV exposure. Computational methods are flexible but expensive—full configuration interaction calculations scale factorially with system size, so they're impractical for anything beyond very small molecules. Semi-empirical methods are fast but inherit whatever errors were baked into their parameterization. DFT sits in the middle: reasonably accurate for many systems, but you can't trust it blindly because the error landscape is uneven. Some functionals work well for main-group elements and fail for transition metals. Others do the opposite. If you need high-accuracy ionization energies for benchmarking, coupled-cluster methods like CCSD(T) are the gold standard, but they're computationally expensive and limited to relatively small systems. For larger molecules, you're usually trading accuracy for feasibility. The practical approach is to run calculations with at least two different functionals or methods and see if they agree. If they diverge significantly, you need to dig into why before trusting either result. Looking up values in the NIST Chemistry WebBook is usually the fastest route if you're just starting out and need reference numbers. It's peer-reviewed and continuously updated. But when you're dealing with a system that isn't in the database—unstable intermediates, exotic oxidation states, or novel compounds—you're on your own and need to fall back on one of the methods described above. That's where understanding the limitations matters more than understanding the definitions.