Getting Equilibrium Data Without Losing Your Mind
When I first started working with equilibrium problems, I kept making the same mistake — dropping the ICE table method entirely for anything that looked messy. That cost me three extra hours on a single batch yield calculation last November, and I've been slightly more careful ever since. The method itself is straightforward enough, but the way you handle it under pressure matters more than most textbooks admit. Most people treat Ice Tables In Chemistry as a rigid fill-in-the-blank exercise. You set up the initial concentrations, changes, and equilibrium values, then solve for x. That's the surface-level version. The practical version involves knowing when to stop using it and switch to something else before you waste an afternoon.
Setting Up Ice Tables In Chemistry Correctly the First Time
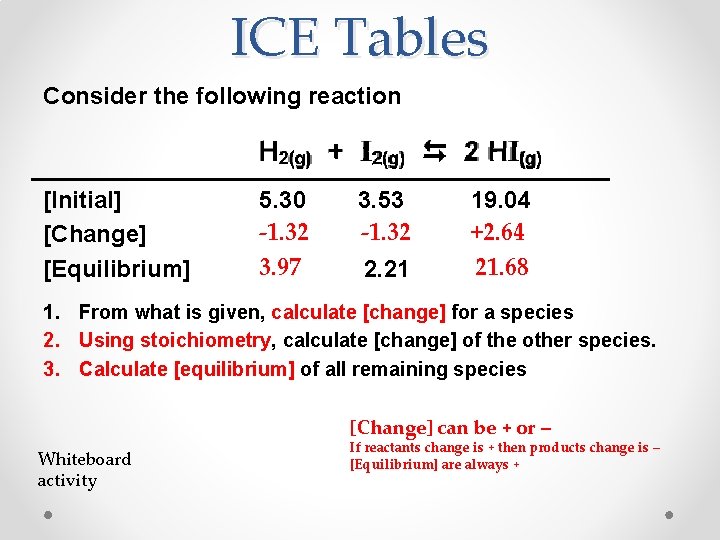
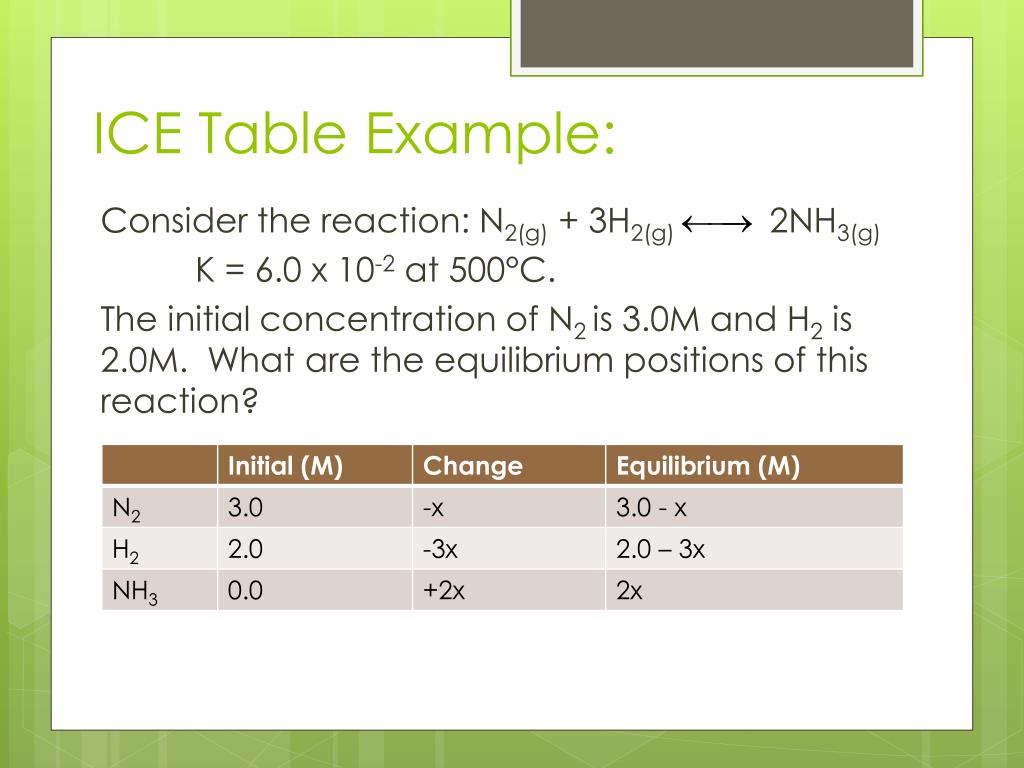
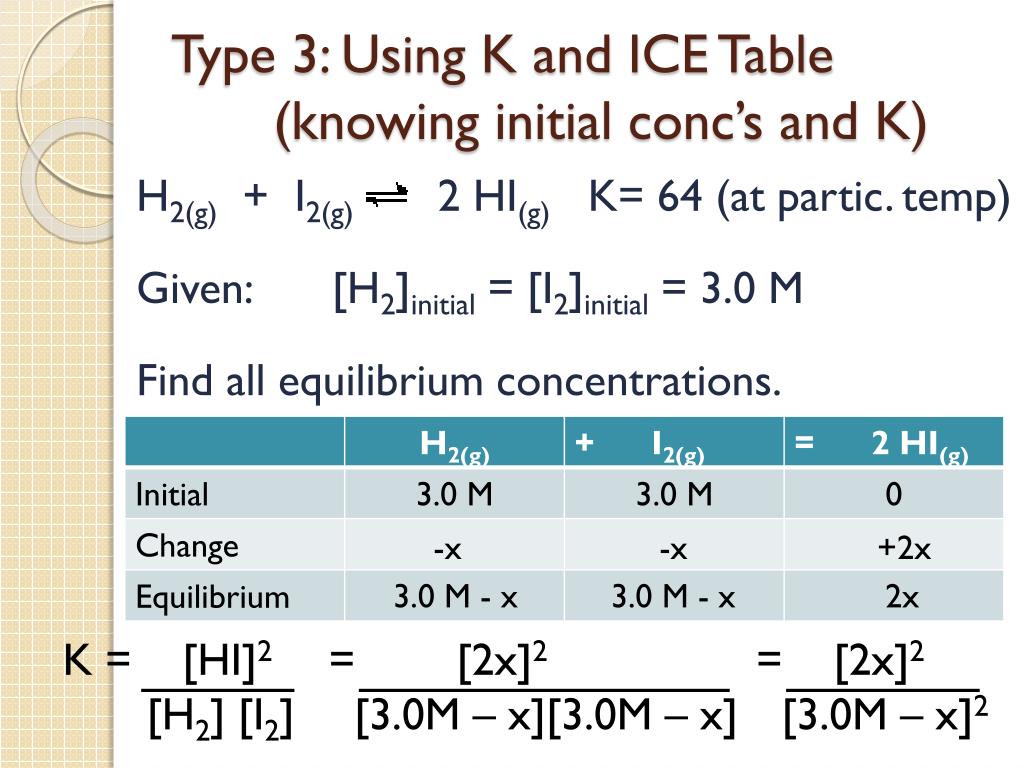
Start with the balanced equation. This sounds obvious, but I've seen analysts skip it because they're confident in their memory of the stoichiometry. Last quarter, a colleague of mine used an ICE table for the decomposition of N2O5 without writing out the full balanced reaction first. He missed that the stoichiometric ratio between N2O5 and O2 was 2:1, not 1:1. His calculated half-life was off by nearly forty percent. The data came back from QA six days later with a single line noting the discrepancy. It would have taken him two minutes to write the equation down. Write the balanced equation. List your initial concentrations or partial pressures. Label them clearly as I, C, and E rows. The change row follows the stoichiometry — if your coefficient is 2, the change term has a 2 in front of x. Don't assume the reader can tell the difference between concentration and partial pressure just from context. Write the units. I know it feels redundant inside a lab notebook, but it saves you when someone else has to review your work three weeks later during an audit. The algebra step is where most people get stuck, and it's usually because they skip the simplification check. If the equilibrium constant is very small relative to your initial concentration — say, Kc is less than 10^-4 and your starting concentration is above 0.1 M — you can approximate that the change is negligible compared to the initial value. This turns a quadratic into a simple division. I do this check every single time before I set up the full quadratic formula. It cuts the calculation from roughly five minutes to under a minute.
When the approximation doesn't hold, use the quadratic formula. There's no shortcut here. I once worked on a synthesis where Kb for ammonia was being used in a buffer system with unusually high ionization. The approximation gave me an answer that was twelve percent off from the exact quadratic solution. That twelve percent translated into a failed stability test for a pharmaceutical intermediate. Regulatory didn't accept it, and we had to redo the entire batch validation. Twelve percent sounds small until it costs you a month of lab time.
Get the Full Details
When Ice Tables Break Down
There are cases where the standard ICE table approach fails outright. The first is any system with coupled equilibria — acid-base plus complex formation, or redox alongside solubility. I ran into this last spring with a mixed chloride-complex system involving silver. Setting up separate ICE tables for each equilibrium created a system of equations that wouldn't resolve cleanly. The workaround was to write a single mass-balance equation that included all species, then solve numerically. I used an iterative solver in Excel rather than doing it by hand. It took about twenty minutes to build the model, and the results matched the literature values within two percent. The second failure case is non-ideal behavior. ICE tables assume ideal solutions where activity coefficients equal one. At ionic strengths above 0.1 M, this assumption starts to introduce systematic error. I've seen this matter significantly in concentrated buffer systems used for HPLC mobile phase optimization. If you're working with samples above 0.1 M ionic strength, consider switching to an activity-corrected model or using a speciation software package instead of hand-calculating. The accuracy gain is worth the learning curve. Another edge case I deal with regularly is gas-phase equilibria where the total pressure changes during the reaction. ICE tables work fine for constant-volume systems, but constant-pressure systems require you to account for the volume change as moles shift. I keep a small reference sheet for this because it comes up frequently in reactor design calculations and I don't trust myself to derive it on the fly anymore. The derivation isn't hard, but it's easy to drop a term when you're tired, and it's late afternoon when I usually do this work.
Practical Workflow That Actually Saves Time
Here's the sequence I use now, and it's mostly about reducing the chance of a silly error rather than anything fancy. Write the balanced equation with states. Note the temperature — equilibrium constants are temperature-dependent, and assuming 25°C when your reaction ran at 60°C will give you wrong answers every time. Look up or calculate K at the actual temperature. Set up the table with concentrations in molarity or pressures in atm, not both in the same table. Check whether the approximation applies. Solve. Plug your result back into the equilibrium expression to verify it satisfies K. If it doesn't, you made an algebra error or the approximation wasn't valid. The verification step is the one most people skip. I've caught two errors this way in the past three months — a sign mistake in the change row and a misread logarithm. Both would have looked plausible until someone tried to reproduce the calculation. Having the check baked into the workflow takes about thirty seconds and prevents that kind of follow-up work. If you need a template, most chemistry department resource pages have downloadable worksheets, and there are also several open-access spreadsheet tools built for this. I keep a personal Google Sheets template with the quadratic solver pre-built so I don't have to reconstruct it each time. It takes about ten seconds to load, enter your K and initial values, and get the result. The manual derivation still matters for understanding, but for routine analysis, the spreadsheet is faster and less error-prone.
The method itself isn't going away. It's a foundational tool for any analytical work involving equilibrium, and it's expected knowledge for anyone working in synthesis, quality control, or process chemistry. The trick is knowing its boundaries — where it simplifies your life and where it hides errors in plain sight.