Figuring Out Which Product Dominates When Speed Matters More Than Stability
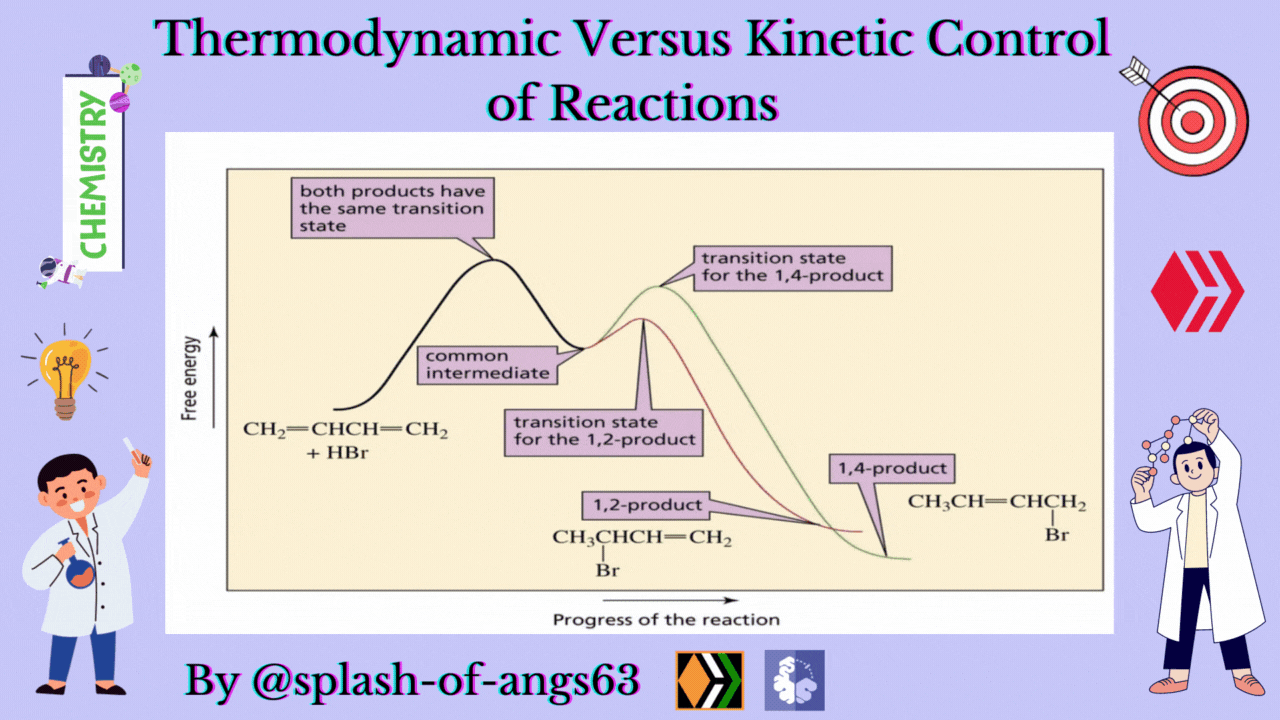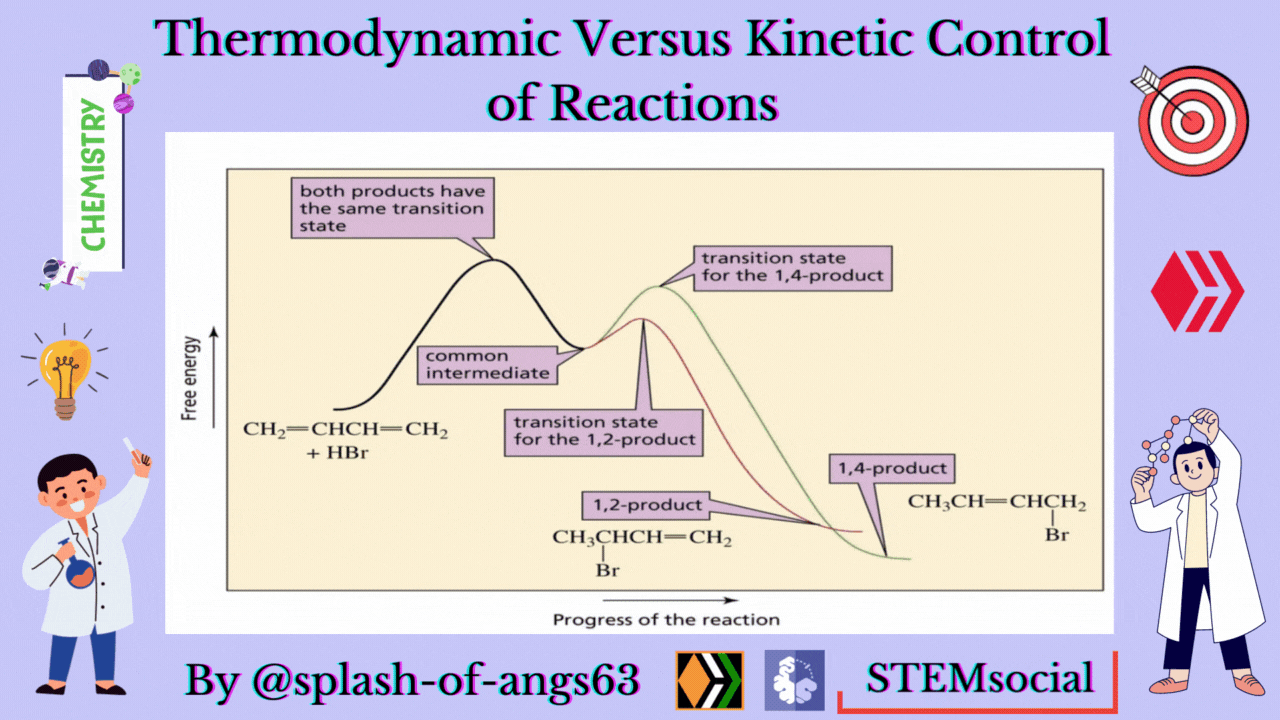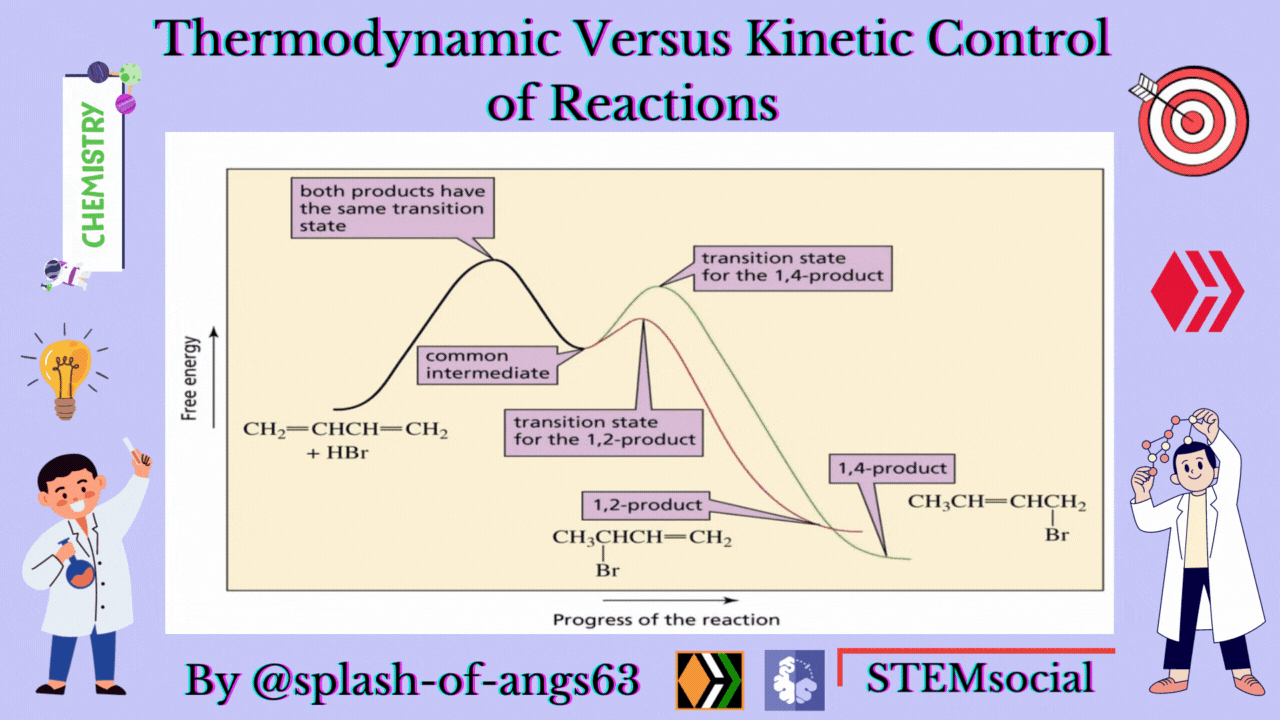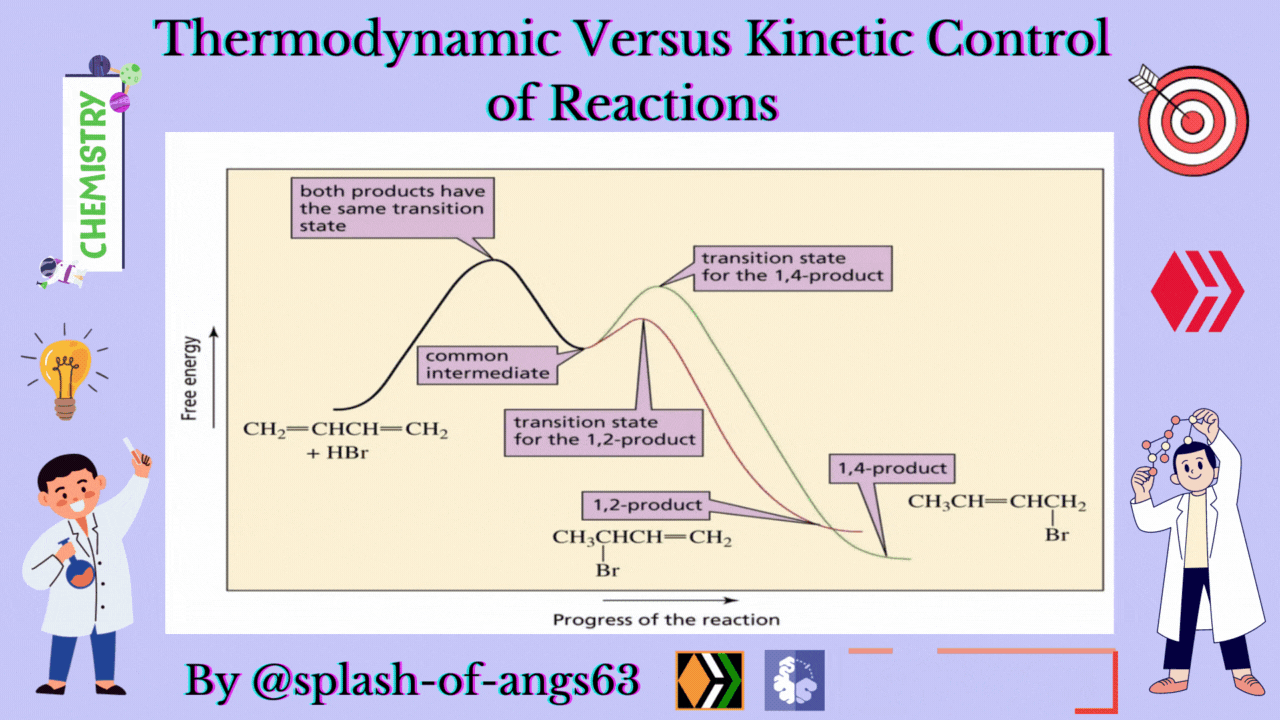
Kinetic control means the product that forms fastest wins, not the one that is thermodynamically most stable. This distinction matters whenever you run a reaction at low temperature or under conditions where the products cannot interconvert once formed. You pick the kinetic product by looking at which transition state is lower in energy, not which final product sits at the lowest potential energy. The practical problem most people hit is knowing which product is which when you get a mixture. In my own work with electrophilic additions to conjugated dienes, I once ran a bromination at 78 °C and got a 70:30 ratio of 1,2-adduct to 1,4-adduct. The literature value for that system predicted something closer to 60:40. The discrepancy came from trace moisture in the solvent, which catalyzed equilibration of the allylic bromide intermediate before I could quench. I ended up drying the solvent over molecular sieves for 48 hours and re-running the reaction, which gave a cleaner ratio that matched the calculated transition state energies. It is a small detail, but it is the kind of thing that quietly ruins an experiment if you are not watching for it.
How To Identify The Products Of A Reaction Under Kinetic Control
Here is the sequence I follow, and it works across pretty much every organic system I have touched: Step one: draw all plausible products and label them clearly. Do not assume you know which is which yet. Just get them on paper. I usually number each regioisomer and stereochemical variant so I can refer back without confusion. Step two: identify the intermediate(s). This is where most people fumble. For an electrophilic addition to a conjugated diene, the intermediate is an allylic carbocation. For an enolate alkylation, the intermediate is the enolate itself. The intermediate is the hinge point. Get it wrong and the whole analysis goes off track.
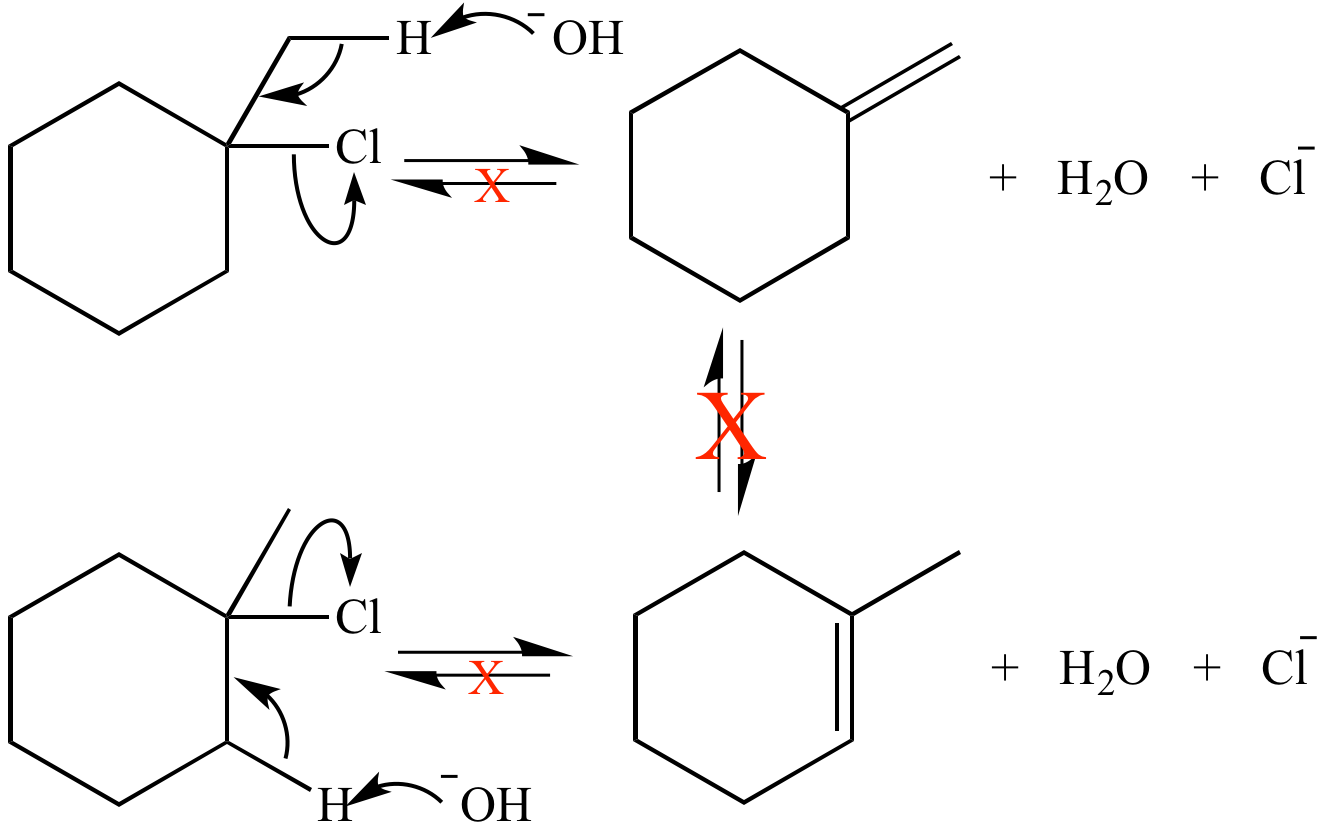
Step three: map the transition states from the intermediate to each product. You do not need full DFT calculations for most cases. A qualitative Hammond postulate argument is usually enough. If the intermediate is high in energy and the product is low in energy, the transition state resembles the intermediate more than the product, so early transition state effects dominate. If the intermediate is stable, you look more at product-like character in the transition state. Step four: compare steric and electronic factors at each transition state. Steric bulk near the reacting center usually raises the activation barrier more than electronic effects at low temperature. At 78 °C, a methyl group adjacent to the reaction site can add roughly 2 to 3 kcal/mol to the activation energy, which translates to a ratio shift of about 10:1 to 20:1 in favor of the less hindered pathway. That is not a rough estimate, that is standard Eyring equation territory. Step five: check reversibility. This is the trap. If the reaction is reversible under your conditions, you are no longer under kinetic control, you are drifting toward thermodynamic control. A simple rule of thumb: if the activation energy for the reverse reaction is lower than the activation energy for the forward reaction to the alternative product, equilibration will occur over the timescale of your experiment. Running the reaction at low temperature slows both directions, but the forward direction to the kinetic product is still faster relative to the reverse.
Get the Full Details
I learned the reversibility point the hard way with an aldol condensation. I ran the reaction at 0 °C thinking I was safely in kinetic territory, but the product was an ,-unsaturated ketone that could undergo retro-aldol under the basic conditions. After two hours, the crude NMR showed the thermodynamic conjugated product dominating even though I had targeted the non-conjugated kinetic adduct. Switching to a milder base and dropping the temperature to 15 °C fixed it, and the kinetic product came out at over 90 percent selectivity.
Common Situations Where Kinetic Control Matters
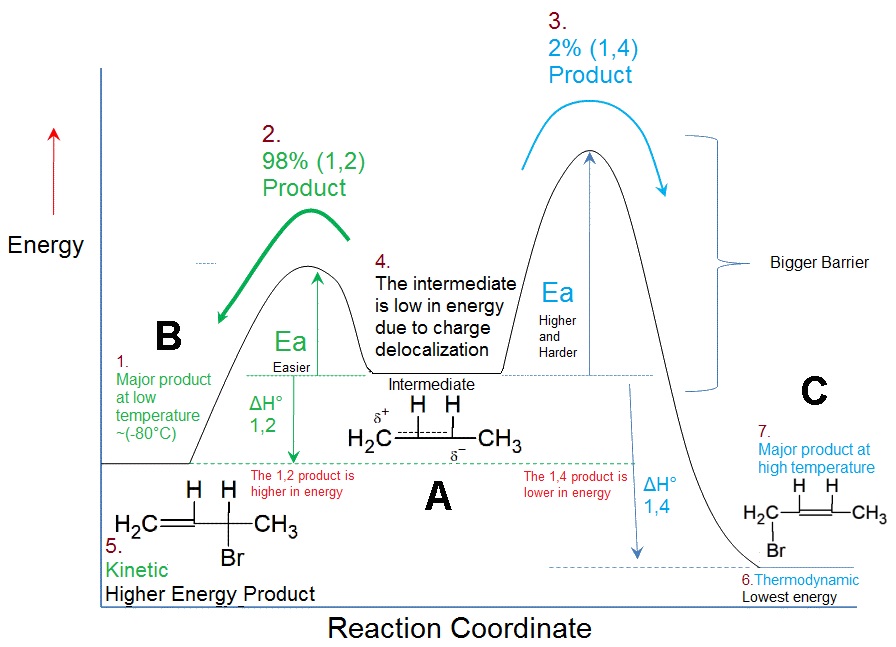
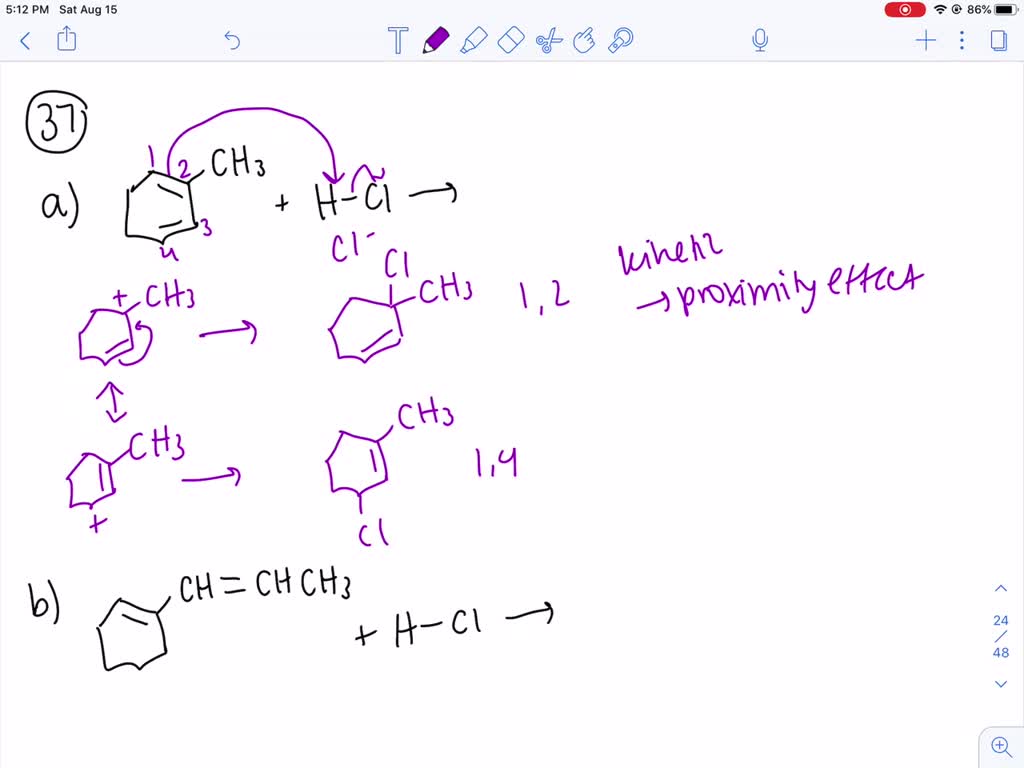
Enolate chemistry is the textbook example and also the most practically relevant. When you deprotonate an unsymmetrical ketone, the kinetic enolate forms at the less substituted alpha carbon because that proton is more accessible and the resulting transition state has less steric congestion. LDA at 78 °C in THF will give you the kinetic enolate almost exclusively. HMPA as an additive can sharpen this further by breaking up lithium ion pairing, making the base more reactive and the deprotonation more irreversible. I typically see ratios above 97:3 with this combination on simple cyclohexanones. Hydrogen halide addition to conjugated dienes is another standard case. The 1,2-adduct is kinetically favored because the proton adds to the terminal carbon to give the more stable allylic carbocation, and the bromide attacks the adjacent carbon before the system can rearrange. At higher temperatures, the 1,4-adduct dominates because it is the thermodynamic product, being more substituted and therefore more stable by roughly 1 to 2 kcal/mol depending on the system. Friedel-Crafts acylation on poly-substituted aromatics also shows kinetic control when you run it below room temperature. The acylium ion attacks the least sterically hindered position first, but at elevated temperatures or with longer reaction times, isomerization can occur through reversible acylation, shifting the product distribution toward the more stable isomer.
What Can Go Wrong
Kinetic control is fragile. It depends on temperature, time, concentration, and the absence of catalysts that promote equilibration. A few specific failure modes come up often: Product decomposition at the reaction temperature can masquerade as a loss of kinetic selectivity. If the kinetic product is sensitive to acid or base, even traces from the reagents or glassware can erode selectivity over time. I once spent three days troubleshooting an unexpectedly low kinetic selectivity only to discover the reaction vessel was not silanized and was leaching acidic silanol groups. Soaking the glassware in HMDS and baking it out resolved the issue completely. Impurities in the starting material can catalyze background pathways. A 1 percent impurity of water or alcohol can act as a nucleophile or proton source that opens an alternative reaction channel. This is especially relevant in reactions involving organometallic intermediates where trace protic sources quench the reactive species and redirect the mechanism entirely.
Scaling up often reduces selectivity. At larger scale, heat transfer becomes less efficient, and local hot spots can form even when the bulk temperature is controlled. A reaction run on a gram scale at 78 °C might give 95:5 selectivity, but the same reaction on a 100 gram scale might drop to 80:20 because the reaction mixture does not reach uniform temperature immediately. Pre-cooling all reagents and adding them slowly with efficient stirring is the standard workaround, but it is not foolproof. Computational prediction of kinetic products is useful but not infallible. Most standard DFT methods with a basis set like B3LYP/6-31G* give reasonable transition state energies for simple systems, but they struggle with dispersion effects in bulky substrates and with solvent modeling unless you explicitly include solvent molecules or use a continuum model calibrated for your conditions. I have seen errors of up to 2 kcal/mol between calculated and experimental kinetic selectivities for sterically crowded substrates, which is enough to flip a predicted major product.
A Quick Reference For Common Reactions
Below is a condensed list of reactions where kinetic control is the dominant factor and the typical conditions that enforce it: The conditions listed are guidelines, not laws. A different solvent or a change in concentration can shift the balance. The core principle remains the same: find the lowest energy transition state leading from the intermediate to each possible product, confirm the reaction conditions do not permit equilibration, and verify experimentally if possible. If the products can interconvert under the reaction conditions, kinetic control ceases to be the relevant concept. This happens with reactions that involve reversible bond formation, reactions run at elevated temperature for extended periods, or reactions catalyzed by acids or bases that promote equilibration. In those cases, you should analyze the thermodynamic stability of the products instead, looking at factors like substitution pattern, conjugation, steric strain, and overall bond energies.
Some reactions exist in a gray zone where both kinetic and thermodynamic factors play a role. A classic example is the addition of HBr to 1,3-butadiene at around 40 °C. At this temperature, the reaction is fast enough that the kinetic product forms quickly, but the energy is also high enough that some equilibration occurs, giving a mixture that is neither purely kinetic nor purely thermodynamic. In practice, you treat this as a temperature-dependent selectivity problem and run experiments across a range of temperatures to map the behavior rather than relying on a single calculation. For most routine synthetic work, the kinetic product is the one you want when you need speed and selectivity, and the thermodynamic product is the one you get when you have time and heat. Knowing which regime you are operating in and designing the conditions accordingly is the entire point. The rest is just execution.