
Understanding Crystal Field Stabilization Energy in Nickel Complexes
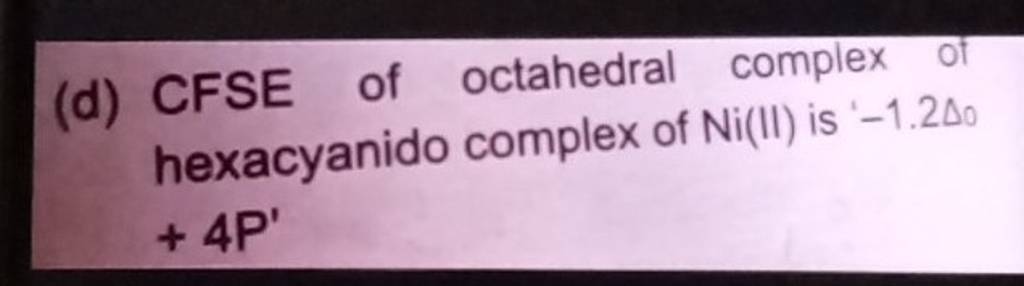
CFSE stands for Crystal Field Stabilization Energy. It is the energy gained when d-electrons occupy lower-energy orbitals in a ligand field compared to a spherical field. For nickel, this gets complicated fast because Ni shows up in several oxidation states and geometries, and the numbers change depending on which one you are looking at. That depends entirely on the oxidation state and geometry. Ni2+ is the most common case and it is also the one people get wrong most often. In an octahedral field, Ni2+ has a d8 configuration. The t2g set holds six electrons and the eg set holds two. The calculation is straightforward: six electrons in t2g contribute -0.4o each, and two in eg contribute +0.6o each. That gives -2.4o + 1.2o = -1.2o. So the CFSE for octahedral Ni2+ is -1.2o, or expressed in Dq terms, -6Dq. In a tetrahedral field the same d8 ion has the orbitals flipped. The e set is lower and the t2 set is higher. Four electrons go into e and four into t2. That works out to -0.6t × 4 plus +0.4t × 4, which equals -0.8t. Since t is roughly four-ninths of o, the tetrahedral CFSE for Ni2+ ends up being only about -0.36o. Much smaller stabilization. This is why tetrahedral Ni2+ complexes are far less common than octahedral ones and why high-spin tetrahedral nickel compounds are almost always paramagnetic with two unpaired electrons.
Now square planar geometry. Ni2+ in a strong field like with CN- ligands goes square planar, dsp2 hybridized. The d-orbital splitting in square planar is messier. The dx2-y2 orbital shoots way up in energy. For d8, all eight electrons pair up in the four lower orbitals and the dx2-y2 stays empty. The CFSE here is significantly larger than octahedral, roughly -2.28o if you use the standard splitting diagram from ligand field theory. That extra stabilization is exactly why [Ni(CN)4]2- is square planar and diamagnetic instead of tetrahedral and paramagnetic. Ni3+ is d7. Octahedral d7 high-spin gives -0.8o. Low-spin d7 gives -1.8o. Ni3+ is rare and usually only stable in specific fluorides or oxides because the third ionization energy is brutal. You will mostly encounter it in solid state materials, not in solution chemistry. Ni0 is d10. Full d-shell. CFSE is zero regardless of geometry. That is why Ni(CO)4 is tetrahedral with no preference driven by crystal field effects at all.
How to Calculate It Step by Step
First, determine the oxidation state of nickel in your complex. Subtract the total charge of the ligands from the overall charge. Ni in NiCl2 is +2. Ni in Ni(CO)4 is 0. Ni in K3[NiF6] would be +3. Second, count the d-electrons. Ni atom is [Ar] 3d8 4s2. Remove electrons from 4s first, then 3d. Ni2+ loses the two 4s electrons to become 3d8. Ni3+ loses two 4s and one 3d to become 3d7. Third, identify the geometry. Octahedral is six-coordinate. Tetrahedral is four-coordinate with no strong field. Square planar is four-coordinate with a strong field ligand. This matters because the orbital splitting diagram changes completely.
Get the Full Details

Fourth, fill the orbitals according to the Aufbau principle and Hund's rule, accounting for pairing energy if the field is strong enough to force low-spin configurations. Then multiply the number of electrons in each set by their energy contribution. t2g is -0.4o, eg is +0.6o in octahedral. e is -0.6t, t2 is +0.4t in tetrahedral. Fifth, sum everything up. The result is your CFSE.
Where People Mess This Up
The most common error is forgetting that CFSE does not include pairing energy. If you are deciding between high-spin and low-spin, you have to add the pairing penalty separately. For Ni2+ octahedral d8, there is no high-spin versus low-spin choice because d8 always fills the same way in octahedral fields. That is a relief. But for d7 Ni3+, you absolutely have to account for pairing energy if you want the real stabilization value. Another error is using o values from one metal and applying them to another. o for [Ni(H2O)6]2+ is roughly 8500 cm-1. o for [Ni(NH3)6]2+ is around 10800 cm-1. These are not interchangeable. If your problem gives you a spectroscopic value, use it. If not, consult the Tanabe-Sugano diagrams for d8, which are well tabulated. I ran into a specific issue once when working with a mixed-ligand Ni2+ complex where one ligand was water and the other was ammonia. The crystal reported an octahedral geometry but the UV-Vis spectrum showed three d-d bands that did not match the standard d8 Tanabe-Sugano prediction for a purely octahedral field. The bands were shifted and split in ways that suggested low symmetry. What was actually happening is the complex had Jahn-Teller distortion, even though d8 is not classically Jahn-Teller active in perfect octahedral geometry. The distortion came from the asymmetric ligand field itself. I resolved it by treating the system as D4h distorted octahedral and recalculating the orbital energies with the split eg and t2g sets. The CFSE changed by about 200 cm-1 from the idealized value, which seemed small but mattered for the thermodynamic prediction I was making.
What CFSE Actually Predicts and What It Does Not
CFSE explains trends in hydration enthalpies across the first-row transition metals. The double-humped curve you see in textbooks, where Mn2+ and Zn2+ are dips and Ni2+ is a peak, is directly attributable to CFSE contributions. For Ni2+, the extra stabilization from its d8 configuration in octahedral fields is why its hydration enthalpy is more negative than you would expect from ionic radius alone. CFSE does not predict whether a complex will form. That depends on lattice energy, solvation energy, entropy, and kinetics. I have seen students treat CFSE as a thermodynamic driving force for complex formation and get confused when their calculated values do not match experimental stability constants. The gap between CFSE and actual free energy of formation can be tens of kilojoules per mole. Ligand field stabilization is real, but it is only one term in the thermodynamic equation. CFSE also breaks down for heavier transition metals where spin-orbit coupling becomes significant. For nickel, which is a 3d metal, this is not a major concern. But if you ever move to Pd or Pt analogs, the whole calculation needs relativistic corrections and you should probably be using a computational method rather than hand-calculating Dq values.
Practical Takeaway
For octahedral Ni2+, memorize -1.2o. For tetrahedral Ni2+, remember it is much weaker at roughly -0.8t. For square planar Ni2+ with strong field ligands, the stabilization is substantially higher and that is what drives the geometry preference. If your assignment or research problem asks for a numerical CFSE value, you need the actual o for your specific complex, usually obtained from absorption spectroscopy or estimated from the spectrochemical series. The spectrochemical series places ligands in order of increasing field strength: I- < Br- < Cl- < F- < OH- < H2O < NH3 < en < CN-
CO. Water gives a weak field, ammonia is intermediate, cyanide is strong. This ordering helps you estimate whether Ni2+ will be high-spin or low-spin, though for d8 octahedral the spin state is always the same regardless of field strength. The geometry is what changes, not the spin multiplicity. If you need to look up o values, the original work by Griffith and Orgel from the late 1950s is still the reference point, though more recent compilations by Lever and by Balhausen are more convenient. There is no downloadable spreadsheet or software that will reliably calculate CFSE for an arbitrary nickel complex without you feeding it the right parameters first. Any tool claiming to do so is either using approximations or pulling from a limited database. The calculation itself is simple arithmetic. The hard part is knowing which o to plug in.


