Working with First-Order Kinetics in the Lab
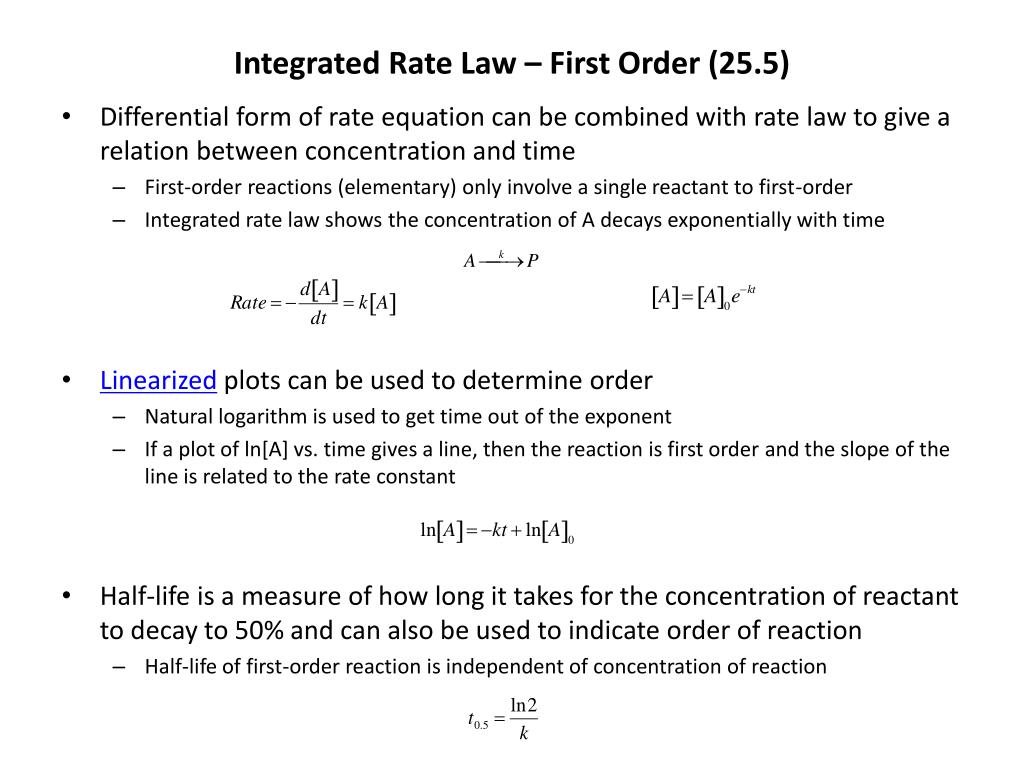
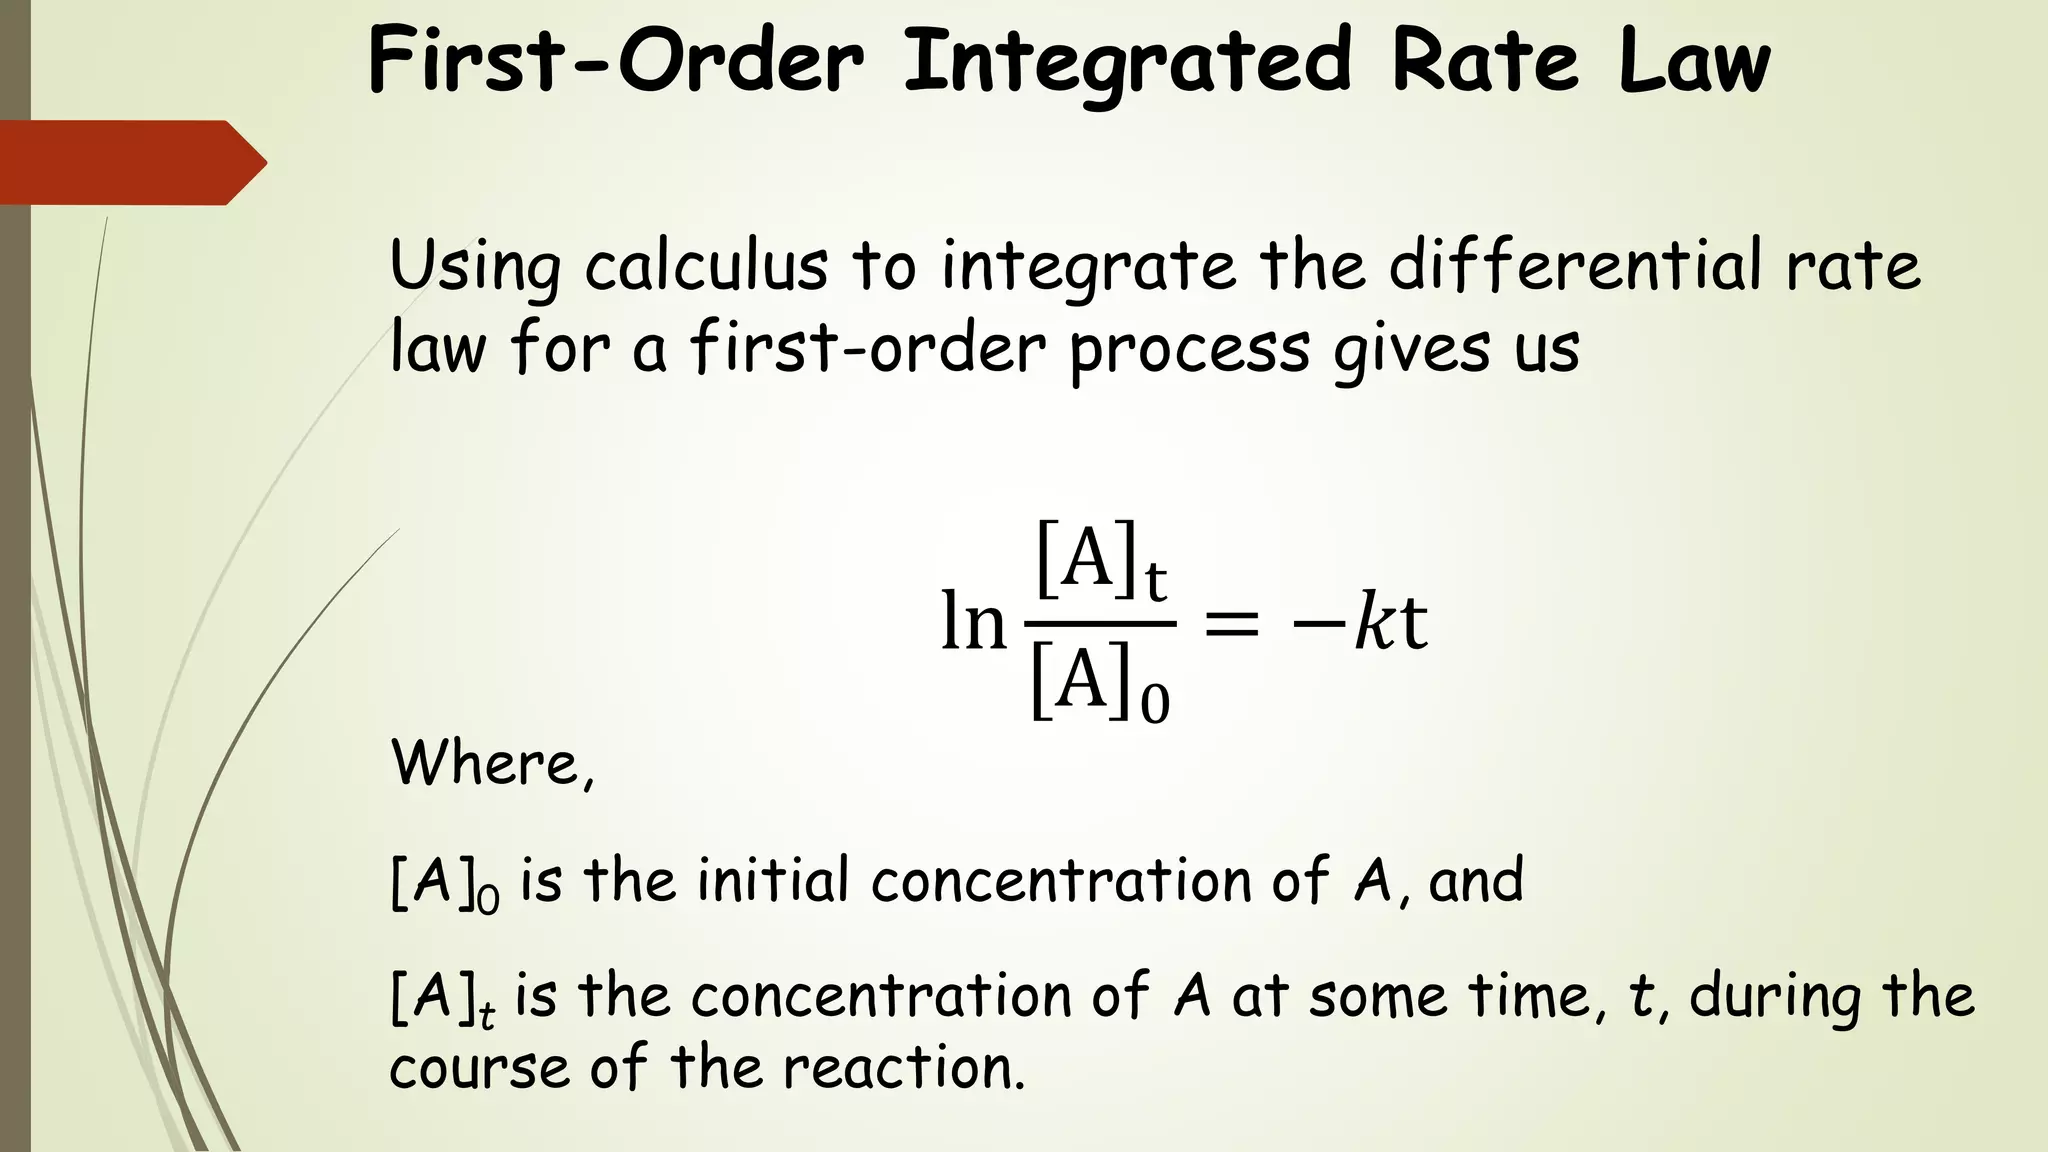
Most people learn the integrated rate law as a equation to memorize for exams. In practice, it's a tool you reach for when you're trying to figure out how long something takes to degrade, whether that's a drug in solution or a contaminant breaking down in groundwater. The math is straightforward. The application is where things get messy. A first-order reaction is one where the rate depends linearly on the concentration of a single reactant. That means if you double the concentration, the rate doubles. The differential form is Rate = k[A], and integrating that gives you the handy form most people actually use: ln[A] = -kt + ln[A]
Or in exponential form: [A] = [A]e^(-kt) k here is the rate constant with units of inverse time, usually per second or per hour depending on your system. [A] is the concentration at time t, and [A] is the starting concentration. The half-life works out to ln(2)/k, which is approximately 0.693/k, and crucially it does not depend on your initial concentration. That is the defining characteristic that separates first-order from second-order or zero-order behavior. I used to think that meant half-life was just a curiosity. It turns out it's the single most practical number you'll work with. If a pharmaceutical company asks you when a compound drops below 90% potency, you don't plug into the full integrated equation. You calculate the half-life, see that 90% is roughly one half-life away, and you have your answer in thirty seconds.
How to Actually Apply It
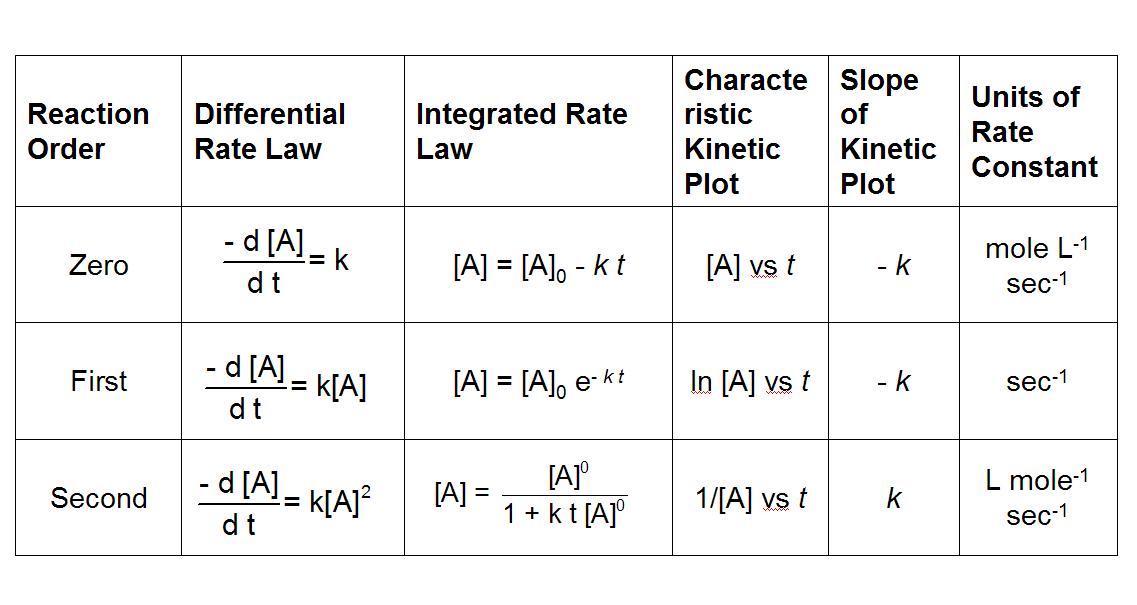
Here's the workflow I use, and it's simpler than what textbooks suggest. You start with experimental data. Collect concentration measurements over time. Plot ln(concentration) versus time. If the plot is a straight line, you've got first-order kinetics. The slope is negative k. The intercept is ln([A]). That's it. That's the whole method. The reason most students get confused is because they try to determine the order by plugging numbers into equations without graphing first. Graph it. A visual check takes two minutes and prevents an hour of wasted work on the wrong model. I had a grad student once spend three days fitting second-order equations to data that was clearly first-order. The residuals were random. The ln plot was a line with an R² of 0.998. He just hadn't looked at the graph.
Get the Full Details

When It Fails You
First-order kinetics assumes a few things that don't always hold. The reaction must be genuinely elementary or pseudo-elementary. If you have a complex mechanism with multiple steps, what looks first-order at low concentration might reveal itself as something else at higher concentration. This comes up constantly in enzyme kinetics and catalysis, but it also shows up in environmental degradation where pH or ionic strength shifts during the reaction. Another practical issue: if your reaction is reversible, the simple integrated form breaks down. You'll see the ln plot curve toward a plateau instead of continuing linearly. I ran into this with a hydrolysis reaction where the product catalyzed the reverse reaction. The first twenty minutes of data looked perfectly first-order. After that, the plot bent. I had to truncate my dataset and fit only the initial linear region, which gave me the forward rate constant. The reverse reaction required a separate experimental setup to characterize. The half-life approach also fails when you're dealing with multi-exponential decay. Some compounds don't degrade in a single step. They go through an intermediate before reaching the final product. The overall decay curve still looks somewhat exponential, but it's actually the sum of two or more first-order processes. Fitting a single line to ln(concentration) versus time will give you an apparent rate constant that changes depending on what time window you choose. If your residuals show a systematic pattern, you're dealing with multi-step kinetics and you need a different model.
Practical Tips That Save Time
Make sure your concentration measurements are in the same units throughout. A common mistake is mixing molarity with mg/L or percent composition without converting. The natural log is dimensionless, but only if all your [A] values use identical units. I've seen reports where the rate constant was off by a factor of one hundred because someone entered ppm for some points and molarity for others. Temperature control matters more than people admit. The rate constant changes exponentially with temperature according to the Arrhenius equation. A variation of just two degrees Celsius can shift k enough to make your linear plot curve slightly. If you're working at room temperature, that means an open lab with windows isn't precise enough. Use a water bath or thermostat. The extra ten minutes of setup saves you from redoing the entire experiment. If you're calculating half-lives for regulatory or compliance purposes, always report the temperature and the method used to determine k. Half-life values float around in literature databases with no conditions attached, which makes them nearly useless for actual predictions. A half-life of six hours means nothing without knowing whether that was measured at 25°C in water or 37°C in buffer at pH 7.4.
When to Use Something Else
Not every reaction is first-order. If your ln plot curves, try plotting 1/[A] versus time for second-order, or [A] versus time for zero-order. The one that gives you a straight line is your model. Sometimes you'll need to fit numerically using software rather than graphing by hand, especially when dealing with reversible or multi-step reactions. Spreadsheet solvers or dedicated kinetic fitting programs like KinTek or COPASI handle this much better than manual calculations, and they give you confidence intervals on your parameters, which you should always report. The integrated rate law for first-order reactions is one of those tools that seems trivial until you need it under pressure. A pharmaceutical stability study, an environmental remediation timeline, a batch reactor optimization. Knowing when it applies, when it breaks, and how to catch the mistakes before they cost you days of work is what separates people who can do the math from people who can actually use the math.