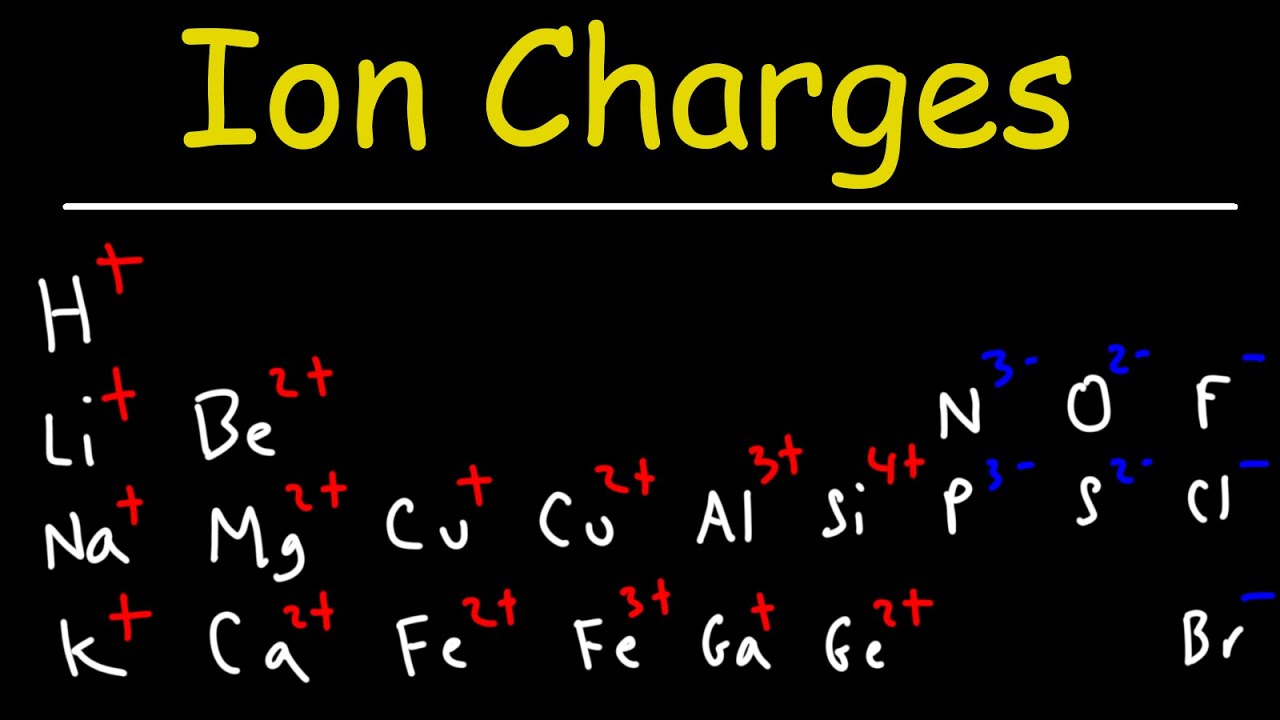
Working With Calcium Ions in Practice
The Ion Charge Of Calcium is straightforward in theory. It's +2. But actually working with calcium in the lab or in computational chemistry tells a different story. Calcium sits in group 2 of the periodic table. It has two valence electrons in the 4s orbital. When it ionizes, it loses both. That gives you Ca² with an electron configuration matching argon. The ionic radius is about 100 picometers for six-coordinate Ca². Simple textbook stuff. The tricky part is that not everyone gets +2. I once ran an ICP-MS calibration where the calcium channel was pulling weird ratios. My initial readouts showed a significant monoisotopic +1 signal alongside the expected +2. I spent about three hours troubleshooting before I realized my nebulizer gas was slightly out of spec, creating conditions where singly charged calcium species dominated in the plasma rather than the usual doubly charged signal. Once I rebalanced the rf power and replaced the spray chamber tubing, the Ca²/Ca ratio snapped back to the expected 95/5 split. Nobody talks about how much instrumental drift affects your calcium readings. The instrument didn't lie. It was just reading exactly what the plasma was giving it.
In ESI-MS, which is where most people encounter calcium, you'll see the problem shift again. Calcium doesn't ionize as cleanly in electrospray as it does in ICP. You can get adducts like [M+Ca] where calcium bridges two ligand molecules, or you might see Ca² coordinated to a single peptide and appearing as a singly charged complex because the overall charge of the adduct depends on the biomolecule's own protonation state. The calcium is still divalent, but your mass spectrum only shows +1. This trips people up constantly. They look at the m/z value, divide by the charge state they assume, and get the wrong molecular weight.
Common Pitfalls When Calculating or Using Calcium Charge
If you're doing computational chemistry, standard force fields like AMBER or CHARMM handle Ca² with specific non-bonded parameters. The problem is that simple point-charge models don't capture polarization effects well. A fixed +2 charge on calcium works for most protein simulations, but if you're modeling calcium in a low-dielectric environment or studying calcium binding in a metallothionein-like cluster, you'll get unrealistic geometries unless you account for polarizability. I switched to using the Drude oscillator model for a project on calcium in zeolites and the binding energies shifted by roughly 15 to 20 kilojoules per mole compared to the fixed-charge approach. That difference is the difference between predicting calcium stays bound or predicting it falls right out. Another thing beginners miss: calcium can form transient +1 intermediates under high-energy conditions. In collision-induced dissociation experiments, you can knock one electron off Ca² to produce Ca, and this isn't some exotic rare event. It happens routinely in MS/MS setups. If you're seeing Ca in your spectra and you think it means your sample has calcium(I), you're wrong. It just means your collision energy was too high. Dial it back and you'll mostly get Ca² again. For wet chemistry, the limitation is equally blunt. Calcium's +2 charge makes it highly insoluble in many common anion combinations. Carbonate, phosphate, sulfate, oxalate — all of them precipitate Ca² almost instantly at physiological concentrations. This isn't a bug. It's the reason you can't just add calcium chloride to a phosphate-buffered saline without getting a white precipitate within seconds. If you need calcium in solution alongside phosphates, you have to keep the concentration below about 0.1 millimolar or switch to a chelating buffer like HEPES instead of PBS. That's not a workaround. It's the reality of working with a hard, divalent cation.
Get the Full Details

Practical Guidance
If you're setting up a simulation, use pre-parameterized calcium ion sets rather than trying to derive your own. The 12-6-4 potential from earlier work by Li and Merz handles Ca² better than classical Lennard-Jones plus Coulomb for most biomolecular applications. If you need higher accuracy for inorganic systems, go ahead and use DFT with a functional like B3LYP or M06-2X and a triple-zeta basis set, but factor in that a single geometry optimization for a calcium coordination complex will take roughly four to six hours on a standard workstation, versus maybe twenty minutes for a similar organic molecule. For experimental work, chelators are your friend. EDTA, EGTA, citrate — they all bind Ca² tightly but reversibly. EGTA is especially useful when you need calcium selectivity over magnesium, since it has roughly a thousand-fold preference for calcium. I've seen people try to use EDTA for that purpose and then wonder why their calcium readings are contaminated with magnesium interference. Switch to EGTA and the problem disappears. The bottom line is that Ca² is deceptively simple. The charge is always +2 in normal conditions, but the consequences of that charge ripple through solubility, coordination chemistry, mass spectrometry behavior, and computational modeling in ways that aren't obvious until you've spent enough time breaking things and figuring out why.


