What Happens When You Actually Try to Measure Solubility
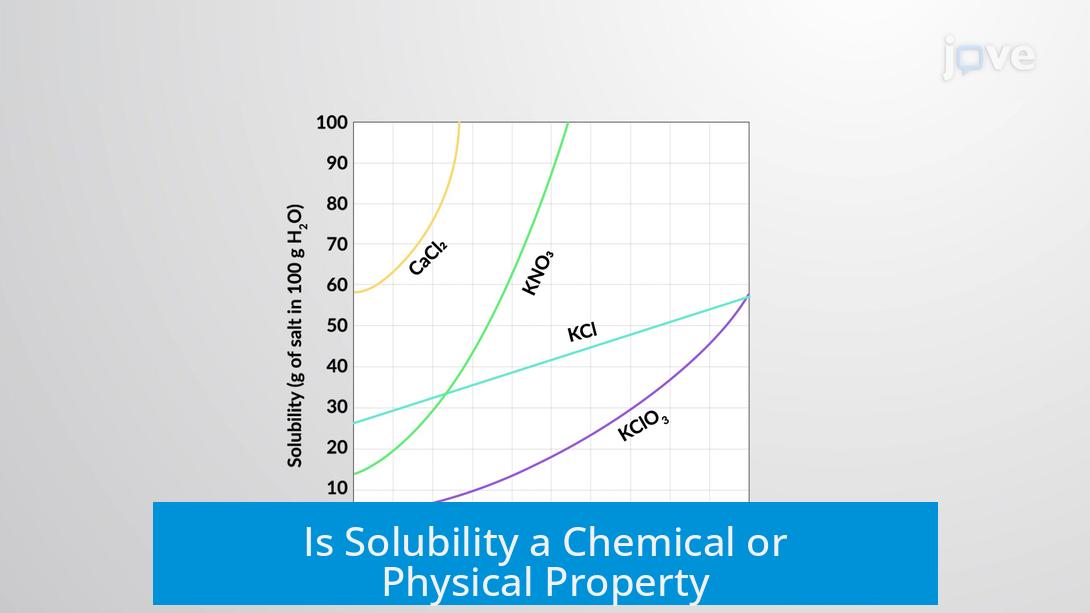
I spent three weeks last year trying to get clean solubility data for a compound that was supposed to be straightforward—sodium chloride in water at various temperatures. The literature values said one thing. My readings said something else entirely. The problem wasn't the method. The problem was that everyone treats solubility as a fixed number you look up and move on from, when it's actually a boundary condition that shifts depending on how dirty your water is, whether your stirring rate was consistent, and whether the container walls had adsorbed impurities from the batch before. This is where the conversation about Is Solubility A Physical Property gets complicated in practice. The textbook answer is yes, obviously. But the person actually running the experiment knows that solubility isn't like density or melting point, where you can measure it once and call it a day. Solubility is a state of equilibrium that takes time to establish, and rushing it gives you wrong numbers every single time.
Is Solubility A Physical Property
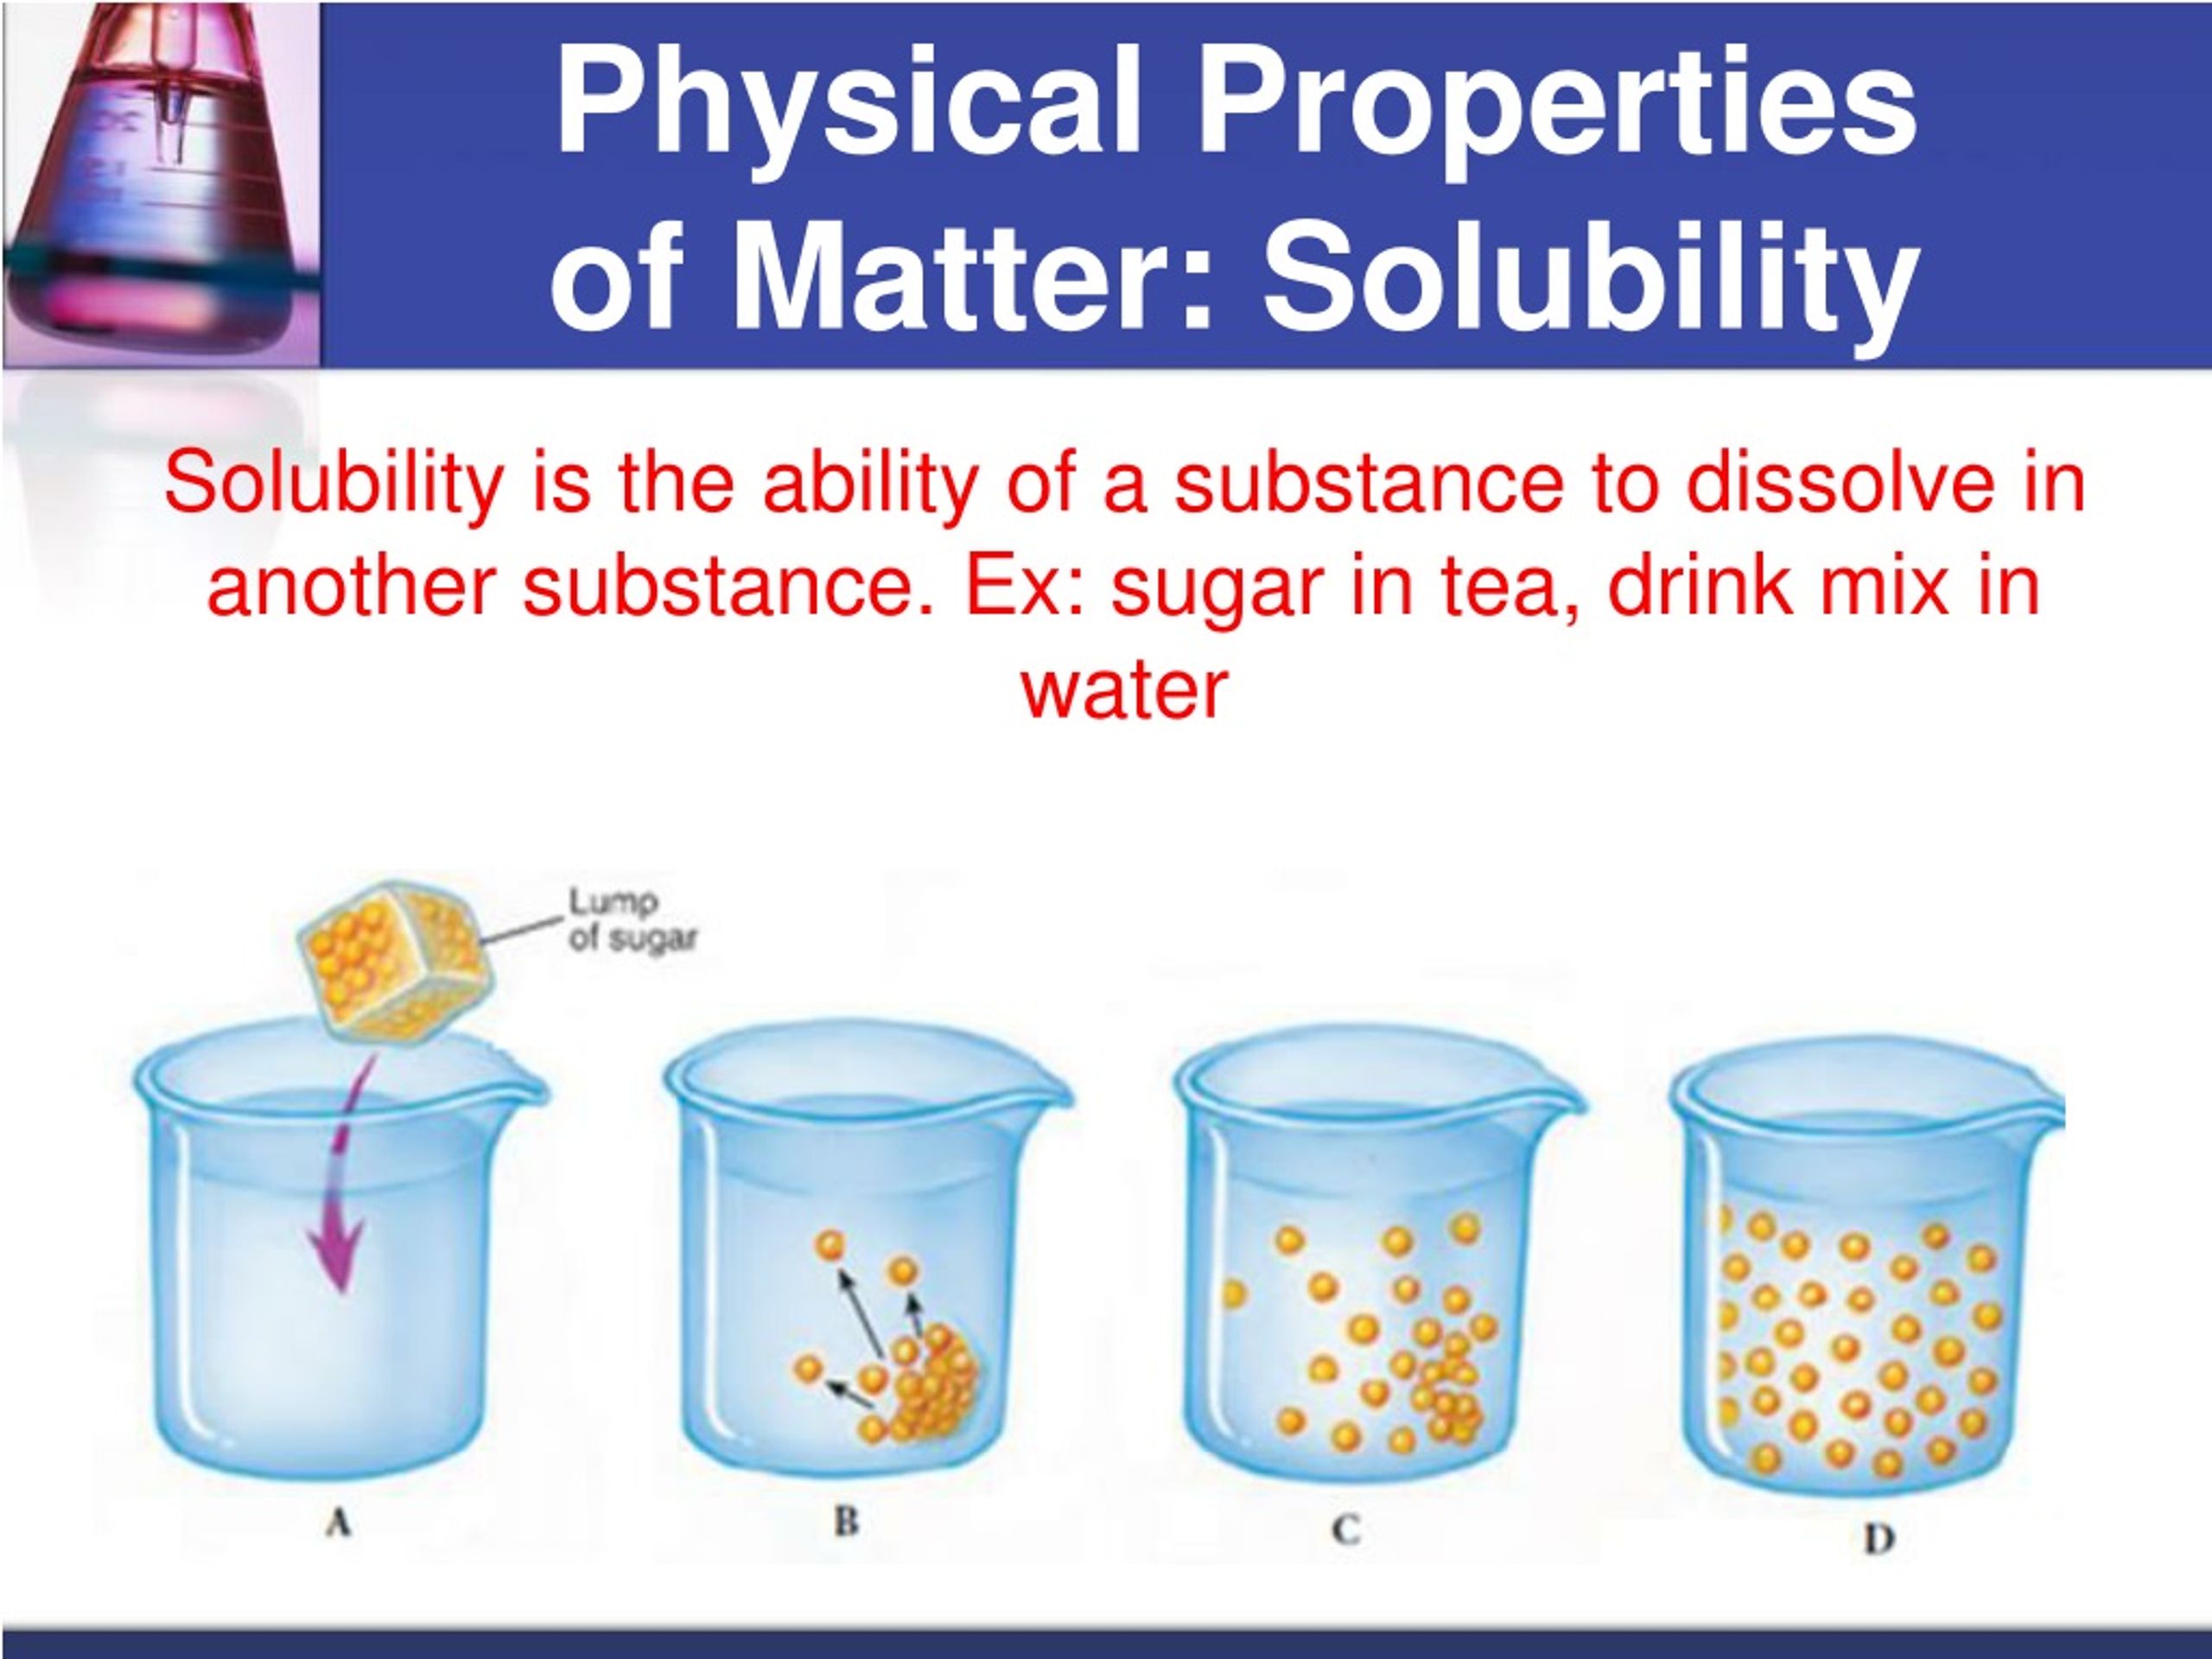
Yes, solubility is classified as a physical property because it describes how a substance behaves under specific conditions without changing its chemical identity. When salt dissolves in water, it's still NaCl. It hasn't become a different compound. You can recover it by evaporation. That's the textbook argument and it's correct for introductory chemistry. The nuance that most people miss is that solubility is also a physical property with some significant caveats. It depends entirely on the solvent system. The same substance has completely different solubilities in water versus ethanol versus hexane. This interdependence on external variables makes solubility feel less like an intrinsic property and more like a system response. That's not wrong—density also depends on temperature and pressure—but the variation in solubility is so dramatically larger that it creates confusion about categorization. Here is another thing that doesn't get explained well. Solubility involves an interfacial process. You're watching a solid-liquid or liquid-liquid boundary reach equilibrium, which means surface area matters. A powdered sample dissolves differently than a single crystal of the same mass, not because the solubility value changes, but because the rate at which equilibrium is reached is completely different. People sometimes mistake kinetic behavior for thermodynamic properties and draw wrong conclusions.
How to Actually Measure Solubility Without Getting Wrong Data
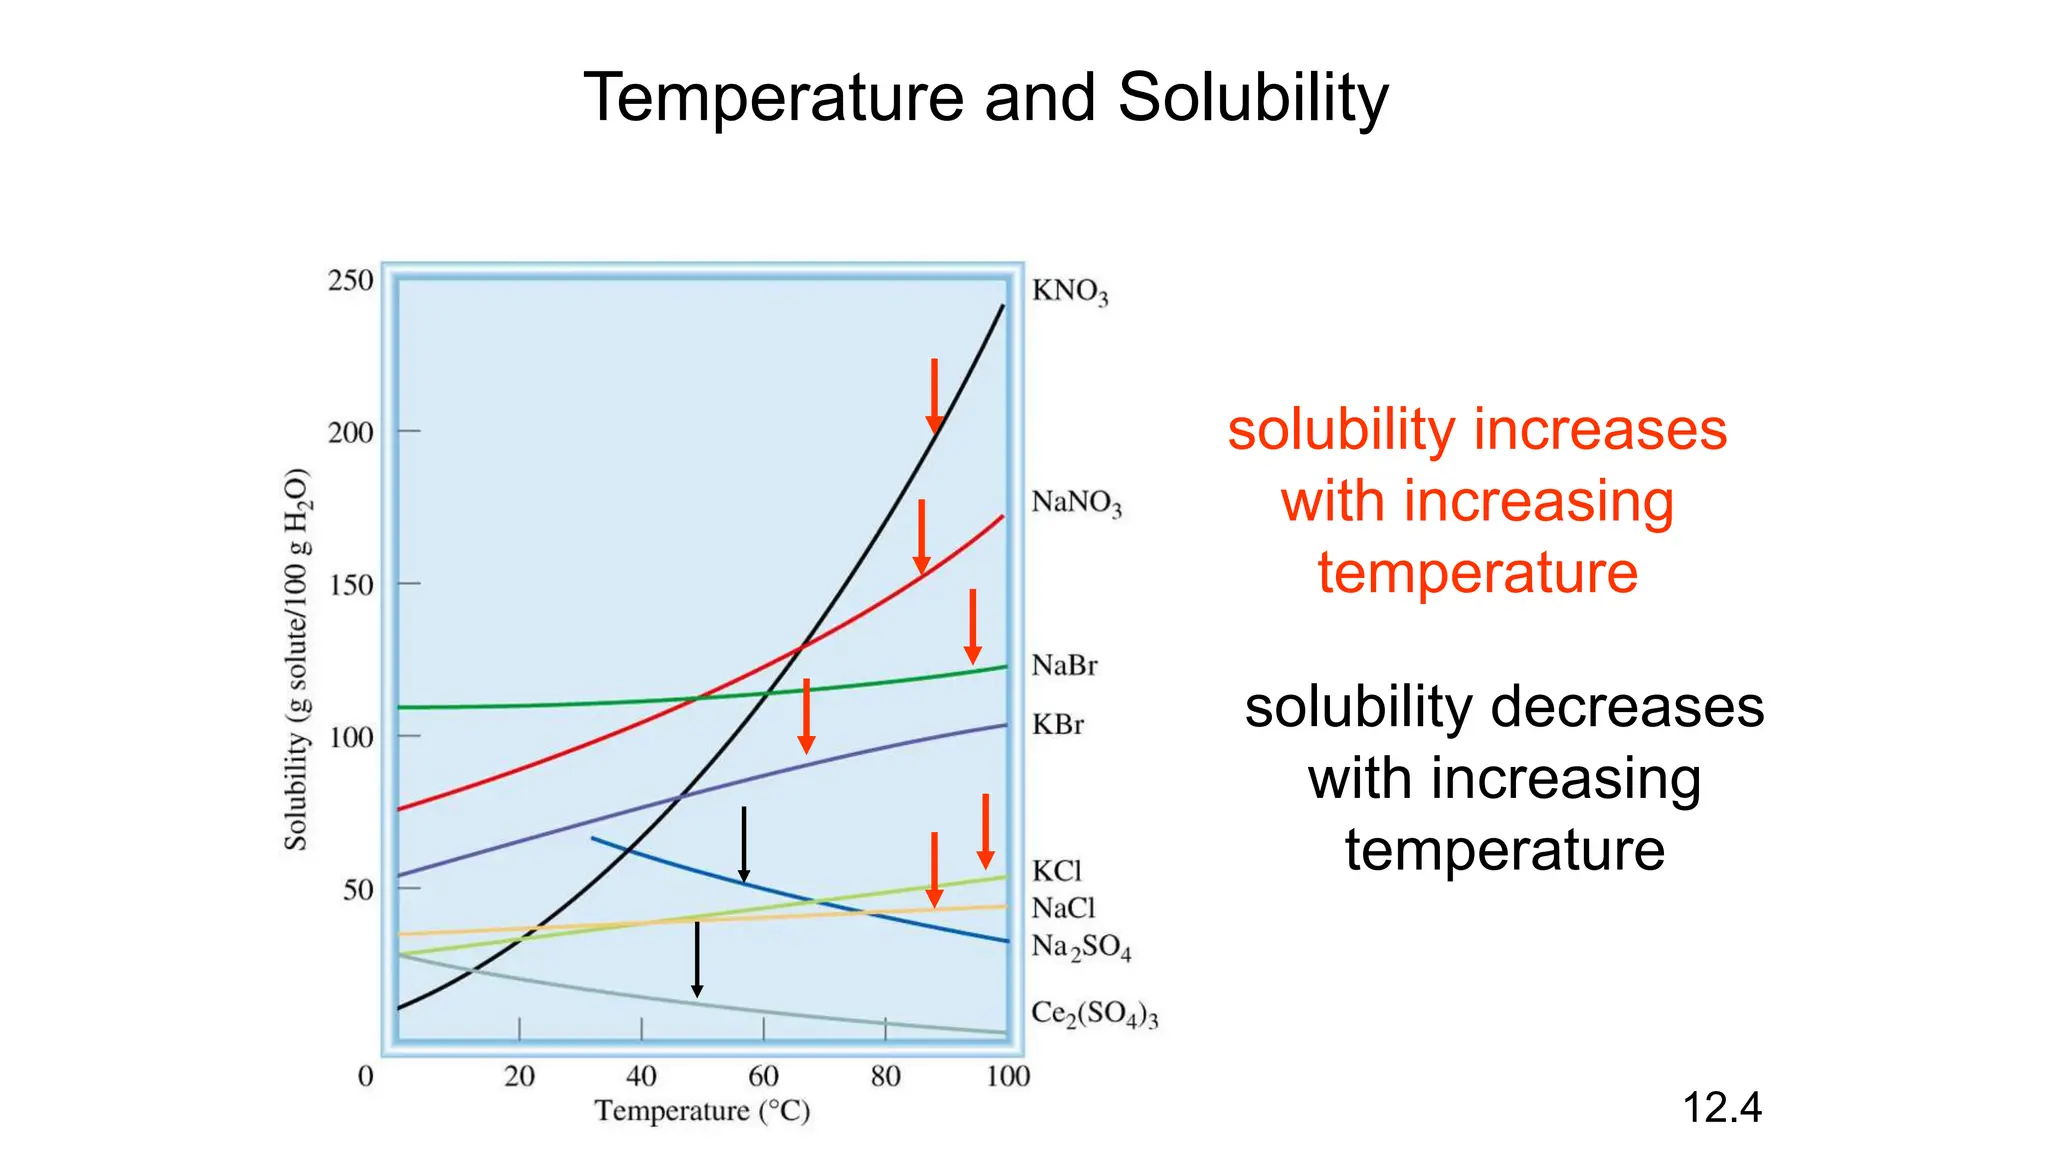
I stopped trying to force solubility data out of quick shake-flask tests and switched to the saturated solution method with extended equilibration times. The procedure is simple enough that you'd think it wouldn't matter, but the execution determines everything. You take an excess of your solute and add it to a known volume of solvent. You seal the container. You stir it at a constant temperature for at least 24 hours, ideally 48. Shorter times leave you with undersaturated samples that you mistake for saturated ones. Then you filter the solution while maintaining the temperature—filtering at room temperature when your experiment was at 60°C will give you precipitate on the filter and a falsely low reading. After that, you take an aliquot of the clear filtrate and analyze it. Gravimetric analysis by evaporating to dryness and weighing the residue is the most reliable approach if your compound is thermally stable. If it decomposes, you use HPLC or titration instead. The problem I ran into with that sodium chloride experiment was that my thermostatic bath had a fluctuation of about 0.5°C. For NaCl this barely matters because its solubility curve is nearly flat. But when I was running the same method on a poorly water-soluble organic compound with a steep temperature dependence, that half-degree swing translated to roughly 8% variance in my measured solubility. I eventually accepted that precision for this type of experiment and reported my values with temperature-controlled uncertainty ranges rather than single numbers.
Get the Full Details
Common Pitfalls That Ruin Your Solubility Measurements
Precipitate formation on filtration. This is the most common error. Your filtrate looks clear and you assume it's a true saturated solution. But if you cooled during filtration, you may have created a supersaturated state or caused premature precipitation that skews your concentration calculation. Ignoring the solvent's purity. I once found that deionized water from our lab's system had trace ammonia contamination that was interfering with the solubility of a weak base I was testing. The water wasn't visibly dirty. The conductivity meter showed normal readings. But the ammonia was enough to shift the equilibrium by creating a common ion effect I never considered. Using freshly distilled water and documenting the solvent source is worth the extra ten minutes. Equilibrium not being reached. Twenty-four hours is a rule of thumb, not a law. For large crystalline samples with low diffusion rates, it can take days. I learned this the hard way with a polymeric solid that I assumed equilibrated in 24 hours. When I ran a second aliquot at 72 hours, the concentration had increased by 12%. The first measurement was wrong and I almost published it.
Polymorphism. This one deserves its own warning. Many organic compounds exist in multiple crystal forms, and each polymorph has a different solubility. The metastable form dissolves faster and gives a higher apparent solubility than the stable form. If you don't know which polymorph you're working with, your data is meaningless for comparison purposes. X-ray powder diffraction before and after the experiment takes five minutes and saves you from having to redo the whole thing.
When Solubility Data Fails You Completely
There are cases where standard solubility measurement is essentially impossible with routine equipment. Amphiphilic compounds that form micelles above a certain concentration create apparent solubility plateaus that aren't real solubility limits—they're just the point where excess compound starts self-assembling instead of dissolving as individual molecules. If you're working with drug candidates or surfactants, you need to distinguish between true molecular solubility and total dissolved material including micellar solubilization. Another scenario where the standard method breaks down is with compounds that undergo chemical degradation in solution. If your substance hydrolyzes, oxidizes, or photodegrades on the timescale of your solubility experiment, you're not measuring solubility. You're measuring whatever decomposition products happen to stay in solution along with your intact compound. Stability-indicating analytical methods are mandatory here, not optional. The workaround I used for a degrading compound was to run the saturation experiment at reduced temperature and then quench the samples immediately with a stabilizing solvent before filtration and analysis. It added steps but eliminated the decomposition artifact that was corrupting my earlier attempts.

Why the Classification Debate Doesn't Actually Matter in the Lab
People argue about whether solubility is a physical or chemical property because they're looking at the definition from the outside. In practice, it doesn't matter whether solubility is technically a physical property. What matters is that you understand the conditions required to measure it reliably and the factors that can invalidate your results. The classification is academic. The measurement is where errors accumulate. My recommendation for anyone setting up solubility experiments is to treat it with the same rigor you'd apply to any quantitative measurement. Document the solvent batch number. Control temperature to within 0.1°C if your compound is temperature-sensitive. Verify equilibrium by sampling at multiple time points. Characterize your starting material for polymorphic form. And don't trust a single data point—replicate everything. The literature values you find online are useful as starting estimates but rarely accurate enough for serious work without verification under your specific conditions. The difference between a published solubility value and your measured value often comes down to solvent purity, particle size, and whether equilibrium was genuinely achieved. Those are controllable variables if you pay attention to them.