Understanding the Trade-Off Between Kinetic and Thermodynamic Control in Organic Reactions
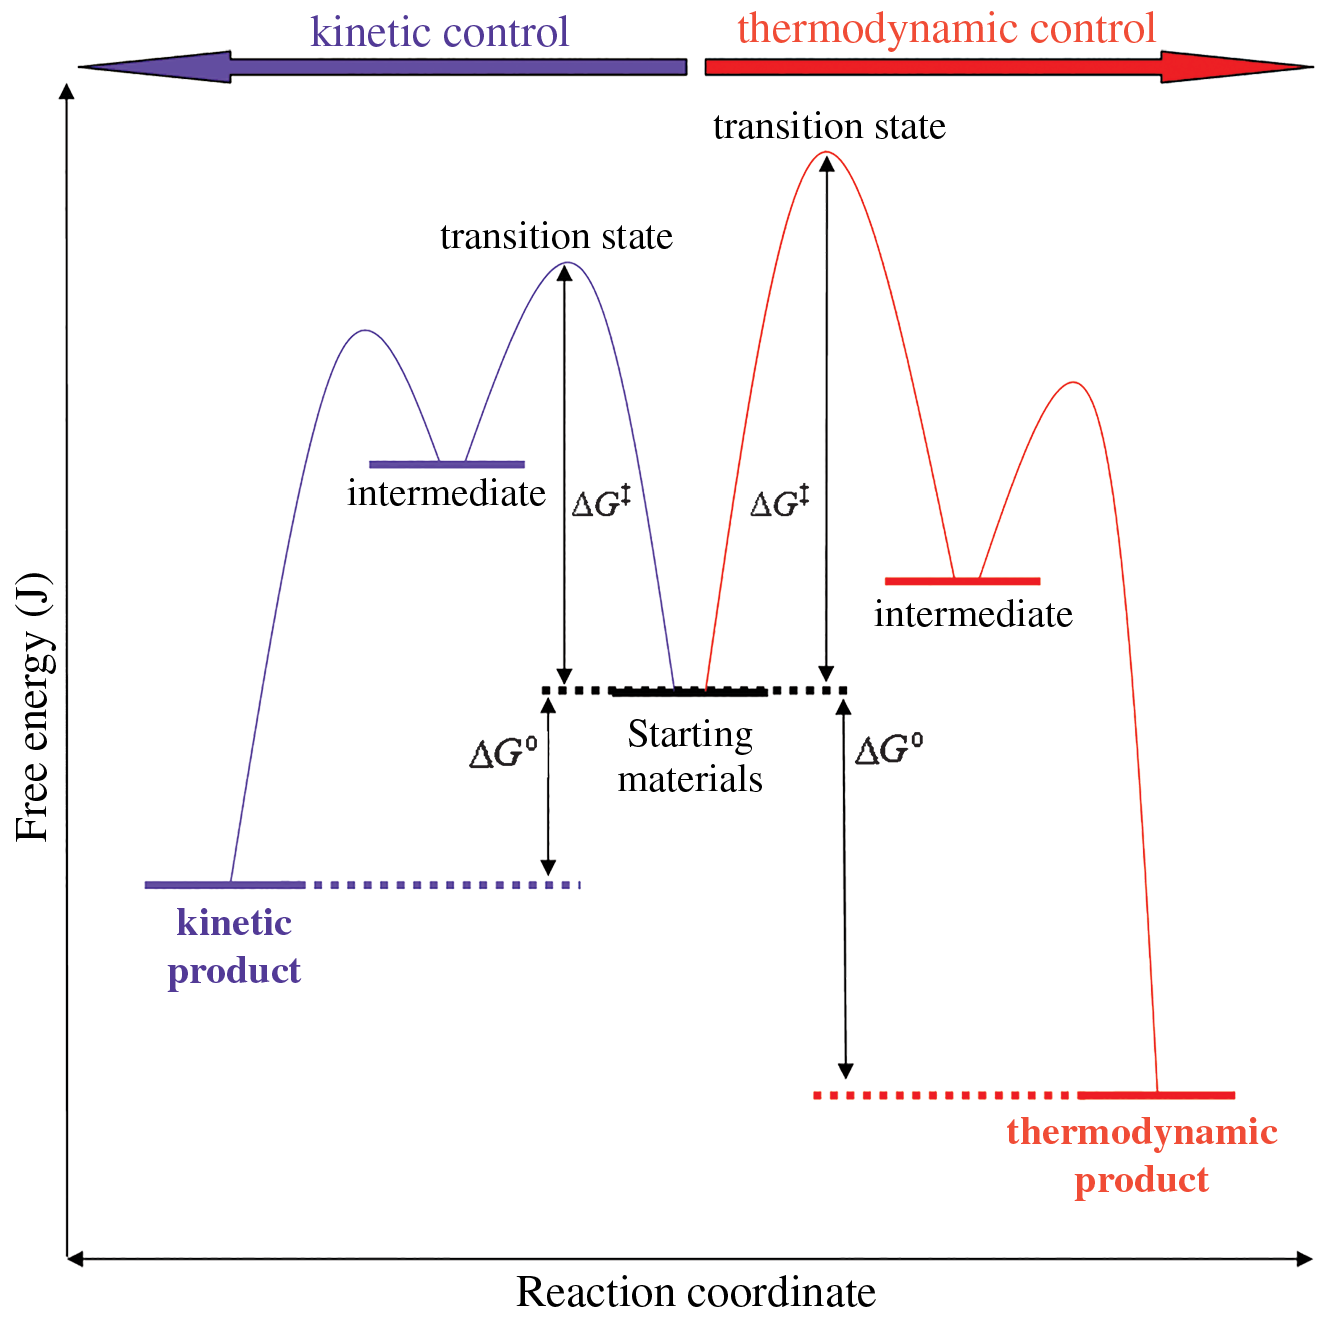
Most undergraduate textbooks present kinetic versus thermodynamic product formation as a neat little dichotomy, and honestly, that works fine if you're just memorizing for an exam. The real world is messier. In practice, you're rarely dealing with a clean switch between one regime and the other. Temperature, solvent, reaction time, concentration, and the presence of catalysts or additives all shift the landscape in ways that standard curriculum glosses over. I've spent years watching chemists trip over the same basic assumptions because nobody bothers to explain what actually happens when you scale something up from a milligram-scale vial to a liter reactor. The core idea isn't complicated. A kinetic product forms faster because it has a lower activation energy barrier. It's the product you get when the reaction is controlled by how quickly things can happen, not by how stable the final result is. A thermodynamic product is the one that sits lower on the energy diagram — more stable overall — but might take longer to form because the path to get there has a higher transition state. Under kinetic control, you quench the reaction early and lock in the fast-formed product. Under thermodynamic control, you let the system equilibrate and the most stable product dominates.
The Practical Guide to Kinetic Vs Thermodynamic Product Selection
Here's how I approach this when I'm actually running reactions. First, I map out the relevant energy landscape. Not the cartoon version you see in textbooks, but the real one. I look at the transition states leading to each possible product and ask which one is actually lower in energy. For enolate-forming reactions, this means I'm thinking about where the deprotonation happens and how reversible it is. For conjugate additions, I'm looking at 1,2 versus 1,4 pathways. The specific reaction type matters less than understanding which bonds are forming and breaking and under what conditions those steps become reversible. Temperature is your primary dial. Lower temperatures favor kinetic control because there isn't enough thermal energy to overcome the higher barrier leading to the thermodynamic product, and more importantly, there isn't enough energy for the products to interconvert once formed. Higher temperatures give the system the freedom to explore both pathways and ultimately settle into the lowest energy state. The rule of thumb is that for every 10 degrees Celsius increase, reaction rates roughly double, but that's a rough guide. The actual relationship is governed by the Arrhenius equation, and the difference in activation energies between your two pathways is what really determines how dramatically temperature shifts the product ratio. Solvent choice is something people underweight. Polar protic solvents can stabilize charged intermediates differently than polar aprotic ones, which shifts relative activation energies. In my experience, switching from ethanol to THF in an aldol-type condensation can flip the major product entirely, not because the thermodynamics changed, but because the solvent stabilizes the transition state for one pathway more than the other. Ion pairing effects in nonpolar solvents can also create selectivities that make no sense when you're thinking purely in terms of energy diagrams drawn in gas phase.
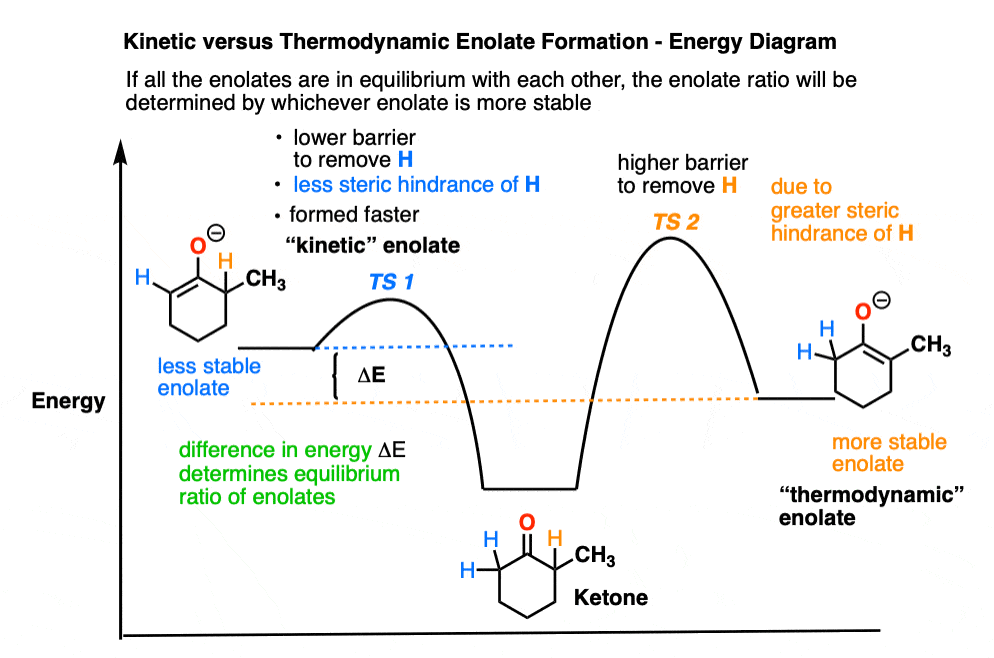
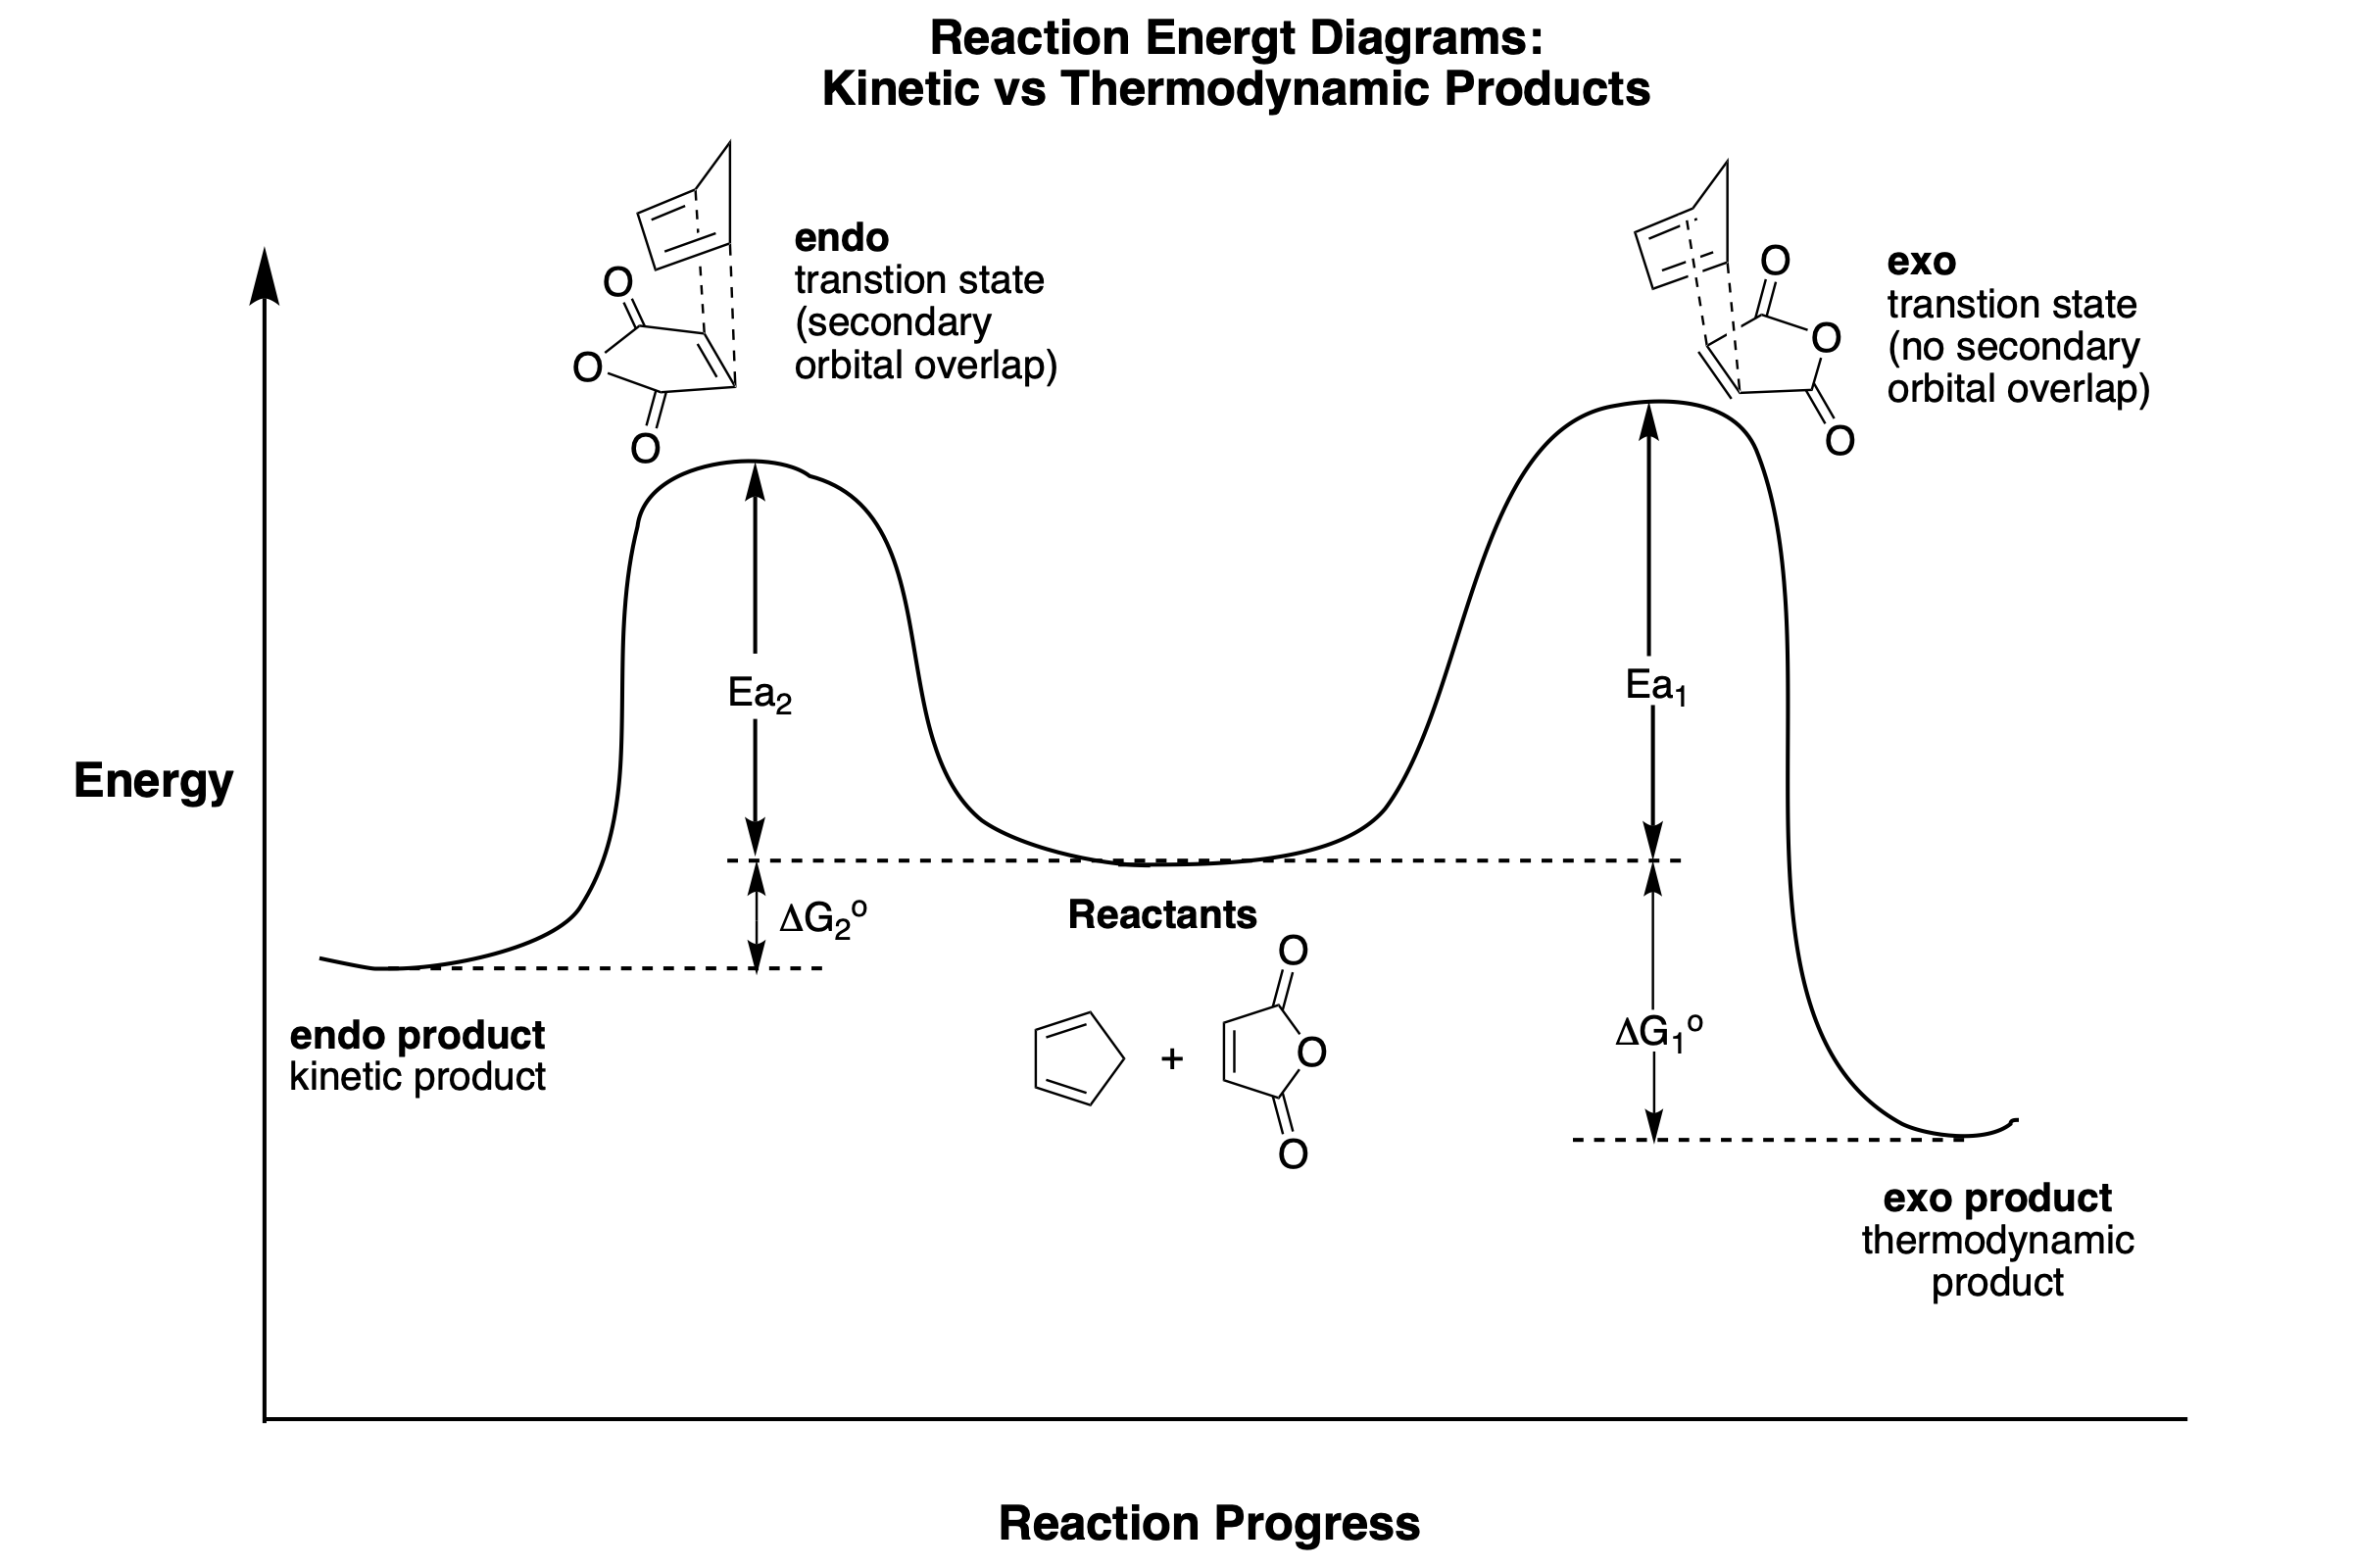
Reaction time is the practical lever. Even if you're running at a temperature where the thermodynamic product is favored, you can isolate the kinetic product if you stop the reaction before equilibrium is reached. This is where things get tricky on scale. In a small vial, heat transfer is nearly instantaneous and mixing is efficient. In a five-liter reactor, temperature gradients of 15 to 20 degrees are not unusual, and mixing times can stretch from seconds to minutes. That means different portions of your reaction mixture are experiencing different conditions simultaneously, and you can end up with a distribution of products that neither condition alone would predict. I learned this the hard way when scaling up a Grignard addition that gave 90 percent kinetic product in a 50 mL flask and about 60-40 mix at the liter scale because the exotherm wasn't being managed properly. For specific reaction classes, the patterns are fairly well established. In enolate chemistry, the kinetic enolate forms with a strong, bulky base like LDA at low temperature in THF, typically around -78 degrees Celsius. The thermodynamic enolate forms with a smaller base like sodium ethoxide at higher temperatures where equilibration is fast. The kinetic enolate has the double bond on the less substituted side because that proton is removed faster — it's more accessible and the resulting transition state is lower in energy. The thermodynamic enolate has the more substituted double bond because that alkene is more stable. Simple enough in theory. In Diels-Alder reactions, the endo product is usually the kinetic product because the secondary orbital interactions stabilize the transition state, while the exo product is often the thermodynamic product because it has less steric strain. But this reverses in certain cases, and I've seen students lose points on exams for not realizing that the endo-exo rule is a tendency, not a law.
Get the Full Details

Electrophilic aromatic substitution follows similar logic. At lower temperatures, the product distribution reflects the relative rates of formation at different positions. At higher temperatures or with longer reaction times, isomerization can occur if the substituents are labile, shifting the mixture toward the more stable isomer. One thing I wish was emphasized more in training is the concept of partial thermodynamic control. Most real reactions exist somewhere in between pure kinetic and pure thermodynamic regimes. The products aren't irreversibly locked in once formed — there's always some back-reaction happening, just at a rate that depends on temperature and the specific bond energies involved. If you're getting a 70-30 mixture and assuming it's purely kinetic, you might be wrong. It could be a partially equilibrated system where the thermodynamic product is still gaining ground. Running a time-course experiment, taking aliquots at different intervals and analyzing them, is the only way to know for sure what's happening. This takes about 30 minutes to 2 hours depending on your analytical method, and it saves you from making incorrect assumptions that could cost you days of follow-up work. Another underappreciated factor is the concentration of reagents. In reactions where the kinetic and thermodynamic pathways have different molecularities — say, a unimolecular rearrangement competing with a bimolecular trap — concentration becomes a third variable alongside temperature and time. Higher concentrations favor the bimolecular path regardless of the energy profile. I encountered this specifically when working with a rearrangement reaction where the kinetic product was formed via an intramolecular shift and the thermodynamic product required intermolecular capture by an external nucleophile. At high substrate concentration, the kinetic rearrangement dominated simply because the molecules found each other faster intramolecularly. Diluting the reaction by a factor of ten flipped the selectivity dramatically. Standard textbooks don't cover this because they usually assume dilute solution conditions.
Now for the limitations. This framework breaks down in several important scenarios. When multiple products have very similar activation energies, temperature shifts produce minimal change in selectivity, and you're essentially gambling. When the kinetic product decomposes or reacts further under the reaction conditions, you might never observe it regardless of how quickly you quench. This is common with reactive intermediates like enolates that can undergo side reactions faster than the desired transformation. When the thermodynamic product is not the most stable possible isomer but merely the most stable among the ones that can actually form through the available pathways — which is almost always the case — you need to be careful about what you're calling "thermodynamic control." Sometimes the reaction simply cannot access the global minimum on the energy landscape due to kinetic barriers that are insurmountable under practical conditions. The biggest practical mistake I see is treating this as a binary choice. In reality, you're often optimizing a complex response surface with temperature, time, solvent, concentration, and additive effects all interacting with each other. A factorial experimental design is usually more efficient than trying to reason through every combination. I also recommend using computational chemistry — even semi-empirical methods like PM6 or DFT at the B3LYP/6-31G level — to estimate relative activation energies before committing to experiments. A single calculation can save you weeks of trial and error, though you should always validate the computational predictions with at least one experimental data point because method choice and solvent modeling can introduce significant errors. If you're working on a synthesis where product selectivity is critical and the kinetic-thermodynamic framework isn't giving you clean results, consider switching reaction mechanisms entirely. Sometimes the most reliable way to control selectivity isn't to tweak temperature and time but to change the protecting groups, use a different coupling reagent, or employ a catalytic system that imposes steric or electronic constraints favoring one pathway. This is what I did when I was trying to achieve consistent selectivity in a multistep synthesis where every temperature adjustment produced marginal gains at best. Switching from a thermal rearrangement to a metal-catalyzed variant gave me 95 percent selectivity where the thermal approach could only manage 70-80 percent even after extensive optimization. The catalytic route added one step to the sequence but eliminated an entire purification bottleneck that was costing me two days per batch.