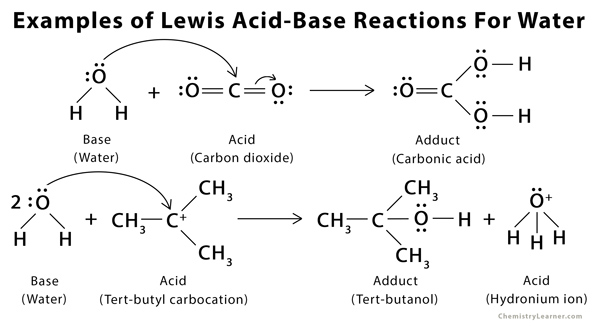
What actually happens when electrons get shared unevenly
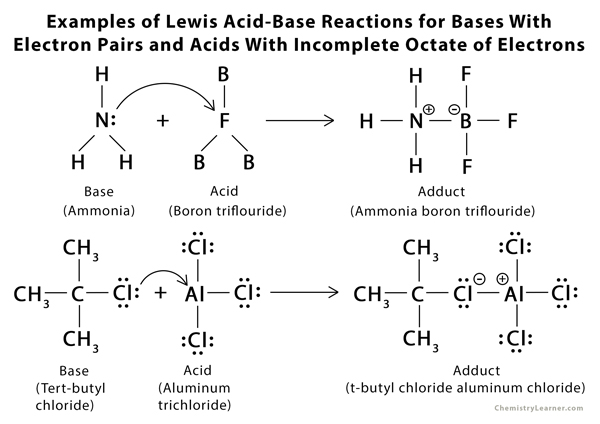
A Lewis acid is an electron pair acceptor. A Lewis base donates an electron pair. That's the definition you'll find in every textbook. The useful part is what those definitions look like on the bench. The most common one people trip over is boron trifluoride. BF sits there with an incomplete octet on the boron. It wants two more electrons. Throw in ammonia and you get FBNH, a clean dative bond. I ran this reaction last year with technical-grade BF in a sealed tube. The gas line was a pain to set up and the adduct precipitated as white needles. Nothing dramatic, just careful handling and a vacuum line that didn't leak. Aluminum chloride does the same thing with chloride donors. AlCl pulls an electron pair from something like diethyl ether and forms a Lewis acid-base complex. That's the basis of Friedel-Crafts acylation, which is still one of the workhorse reactions in organic synthesis. The catch is that AlCl also coordinates to the product oxygen after the reaction finishes, so you need excess reagent and a thorough aqueous workup to liberate the ketone. If you skip the extra equivalent, your yield drops to around sixty percent or worse.
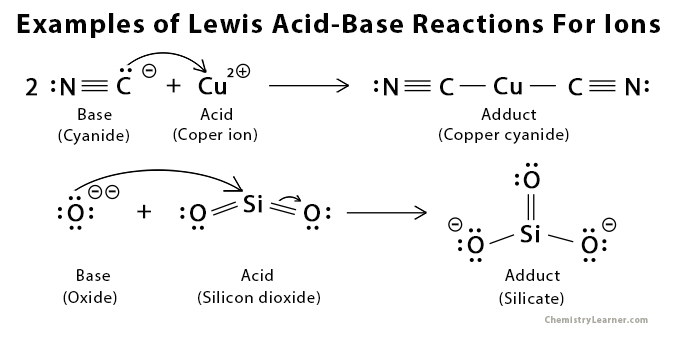
Transition metal ions are another category that gets overlooked. Fe³, Cu², Zn² — they all act as Lewis acids because they have empty d-orbitals that can accept electron pairs. Ligands like water, ammonia, or cyanide are the bases. In practice this shows up in coordination chemistry labs when you're making complexes. The color changes are immediate and useful for identifying what you've got. A blue solution turning deep purple when you add excess cyanide to copper sulfate is a pretty obvious signal. Now for the counter-intuitive bit. Not everything you'd expect to be a Lewis acid actually behaves like one under normal conditions. Silicon tetrafluoride is a classic example. SiF should readily accept electron pairs because silicon can expand its coordination sphere, but the kinetics are slow without a catalyst or elevated temperature. I found this out the hard way when I tried to use it as a fluorinating agent in a mild condition protocol. The reaction barely moved at room temperature. Heating to 80°C with a phase transfer catalyst made it work, but the selectivity went downhill fast. Sulfur trioxide is another tricky case. It's a strong Lewis acid in the gas phase, but in solution it tends to self-associate into trimers and tetramers, which reduces its reactivity toward external bases. If you're doing sulfonation with SO, you usually dissolve it in sulfuric acid first to break up those oligomers. Otherwise you get inconsistent results and a messy product mixture.
On the basic side, amines are the go-to Lewis bases in most synthetic work. Pyridine, triethylamine, DABCO — they all have lone pairs ready to donate. Phosphines are stronger bases but also more air-sensitive. Trimethylphosphine oxidizes rapidly in air, so if you're using it you need to work under inert atmosphere and distill it fresh. I spent a week troubleshooting a cross-coupling reaction before realizing the PMe had turned into phosphine oxide. The catalyst wasn't dead, the ligand was. That cost me about four days of wasted effort. Harden soft theory matters here too. Hard acids like Al³ prefer hard bases like fluoride or oxide. Soft acids like Ag or Pd² prefer soft bases like phosphines or thioethers. This isn't just academic — it determines which reagents you should pair together. Mixing a hard acid with a soft base often gives weak or unstable complexes. You'll see this in catalysis when the wrong ligand causes the metal center to precipitate out of solution instead of staying active. One practical limitation worth noting: Lewis acid-base chemistry doesn't always play nice with protic solvents. Water and alcohols are Lewis bases themselves and will coordinate to your acid reagent, often deactivating it. That's why many Lewis acid-catalyzed reactions run in dry dichloromethane or acetonitrile. If your substrate is moisture-sensitive, you also need to dry the solvent properly. Molecular sieves or distillation over sodium/benzophenone are standard approaches.
Get the Full Details
Cryptic edge case that came up recently: some carboxylic acids can act as Lewis acids through the carbonyl oxygen, but they're much weaker than metal-centered acids. The difference is subtle in mechanism but obvious in outcome. A carboxylic acid won't promote Friedel-Crafts reactions the way AlCl does, no matter how much you use. Trying to substitute one for the other is a common beginner mistake that wastes reagents and time. The reality of working with these systems is that the textbook examples are simple, but real reactions involve impurities, solvent effects, and competing coordination pathways. A good rule of thumb is to test the acid-base interaction on a small scale before committing to a full reaction. A few milligrams in an NMR tube with deuterated solvent will tell you whether complexation is happening and how tight it is. That saves more time than any amount of literature searching.