Drawing the Lewis Structure For Carbonate Ion Step by Step
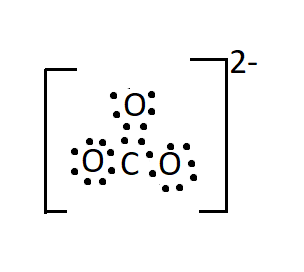
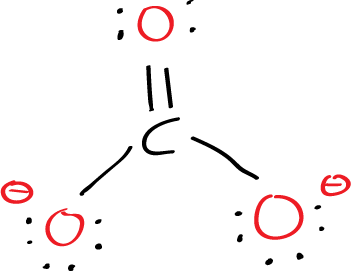
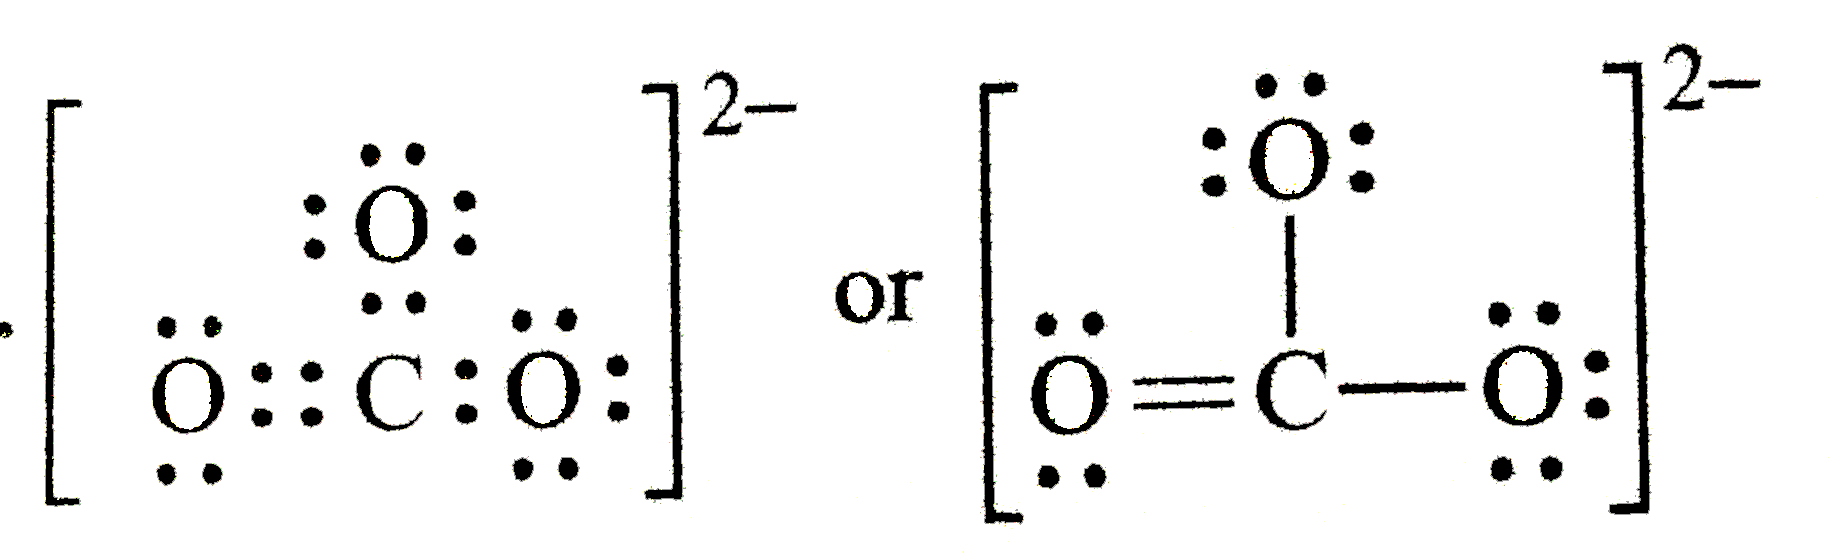
Most people get tripped up on the carbonate ion because they treat it like a straightforward covalent molecule and miss the resonance aspect entirely. The quick version: CO3 2- has one carbon bonded to three oxygens, two of which carry a negative formal charge in any single resonance form, but the real structure is a hybrid where the double bond rotates between all three positions. Start by counting valence electrons. Carbon gives you four, each oxygen gives six (times three = eighteen), and the 2- charge adds two more electrons. That is twenty-four total valence electrons to work with. Put carbon in the center since it is the least electronegative atom, then draw single bonds to each oxygen. That uses six electrons, leaving eighteen. Fill the octets on the three oxygens first, which consumes another eighteen. You are now at twenty-four electrons with zero left over, but carbon only has six electrons from those three single bonds, so it is incomplete. Take one lone pair from one of the oxygens and convert it into a double bond with carbon. That gives carbon an octet and resolves the electron count. The catch is that any of the three oxygens could be the one donating the lone pair, which means you have three equivalent resonance structures, not one definitive drawing. In practice, when I was grading undergraduate lab reports on this, I saw students draw a single Lewis Structure For Carbonate Ion with a fixed double bond on one oxygen and then mark the other two as single-bonded with full negative charges. That is technically one valid resonance contributor, but writing it as the only structure implied a static geometry that does not exist. The workaround I started requiring was simple: draw all three resonance forms with double-headed arrows between them, and add the curly bracket notation showing the overall 2- charge. Students who skipped this step consistently lost points on questions about bond length, because they could not explain why all three C-O bonds in the actual ion measure at 128.8 picometers, intermediate between a typical single bond (around 143 pm) and a double bond (around 120 pm).
Here is something most textbooks gloss over quietly. Formal charge alone does not tell the whole story here. If you assign formal charges in the standard way across any single resonance contributor, the carbon is zero, the double-bonded oxygen is zero, and the two single-bonded oxygens are each negative one. But experimentally, each oxygen carries about a negative two-thirds charge, not a full minus one or zero. The delocalization is spread across all three C-O interactions equally. When you move into computational chemistry or X-ray diffraction work, relying on a single Lewis structure will mislead you about charge distribution and reactivity patterns, particularly at the oxygen sites where nucleophiles or protons would actually attack. Another thing that comes up regularly and causes confusion: the carbonate ion is planar with sp2 hybridization on the carbon, and the remaining p orbital participates in pi delocalization across all three oxygens. This is why the bond angles are exactly 120 degrees, not some distorted variation. If you ever see a structure drawn with bent geometry around the carbon, that is wrong, and it usually stems from someone forgetting that the double bond does not lock into one position. The limitation you need to accept is that Lewis structures are inherently crude for ions like carbonate. They show connectivity and approximate electron accounting, but they cannot represent the actual electron density distribution. Quantum mechanical calculations or even molecular orbital diagrams do that job. For most general chemistry contexts, three resonance forms with proper charge notation is the standard acceptable answer, but if you are working in inorganic synthesis or studying reaction mechanisms involving carbonate as a ligand or base, you should think in terms of delocalized pi systems rather than flipping between static drawings. The model breaks down quickly when you try to use it for quantitative predictions about acidity, binding affinity, or vibrational spectra.