What You Actually Need to Know About Linkage Mapping
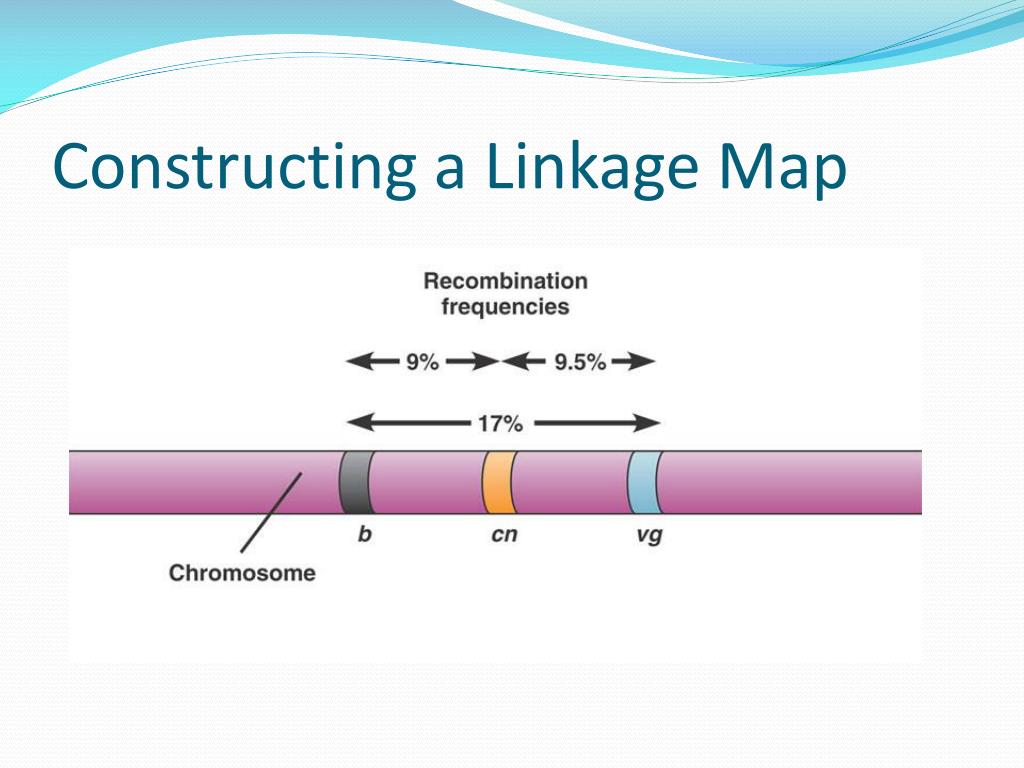
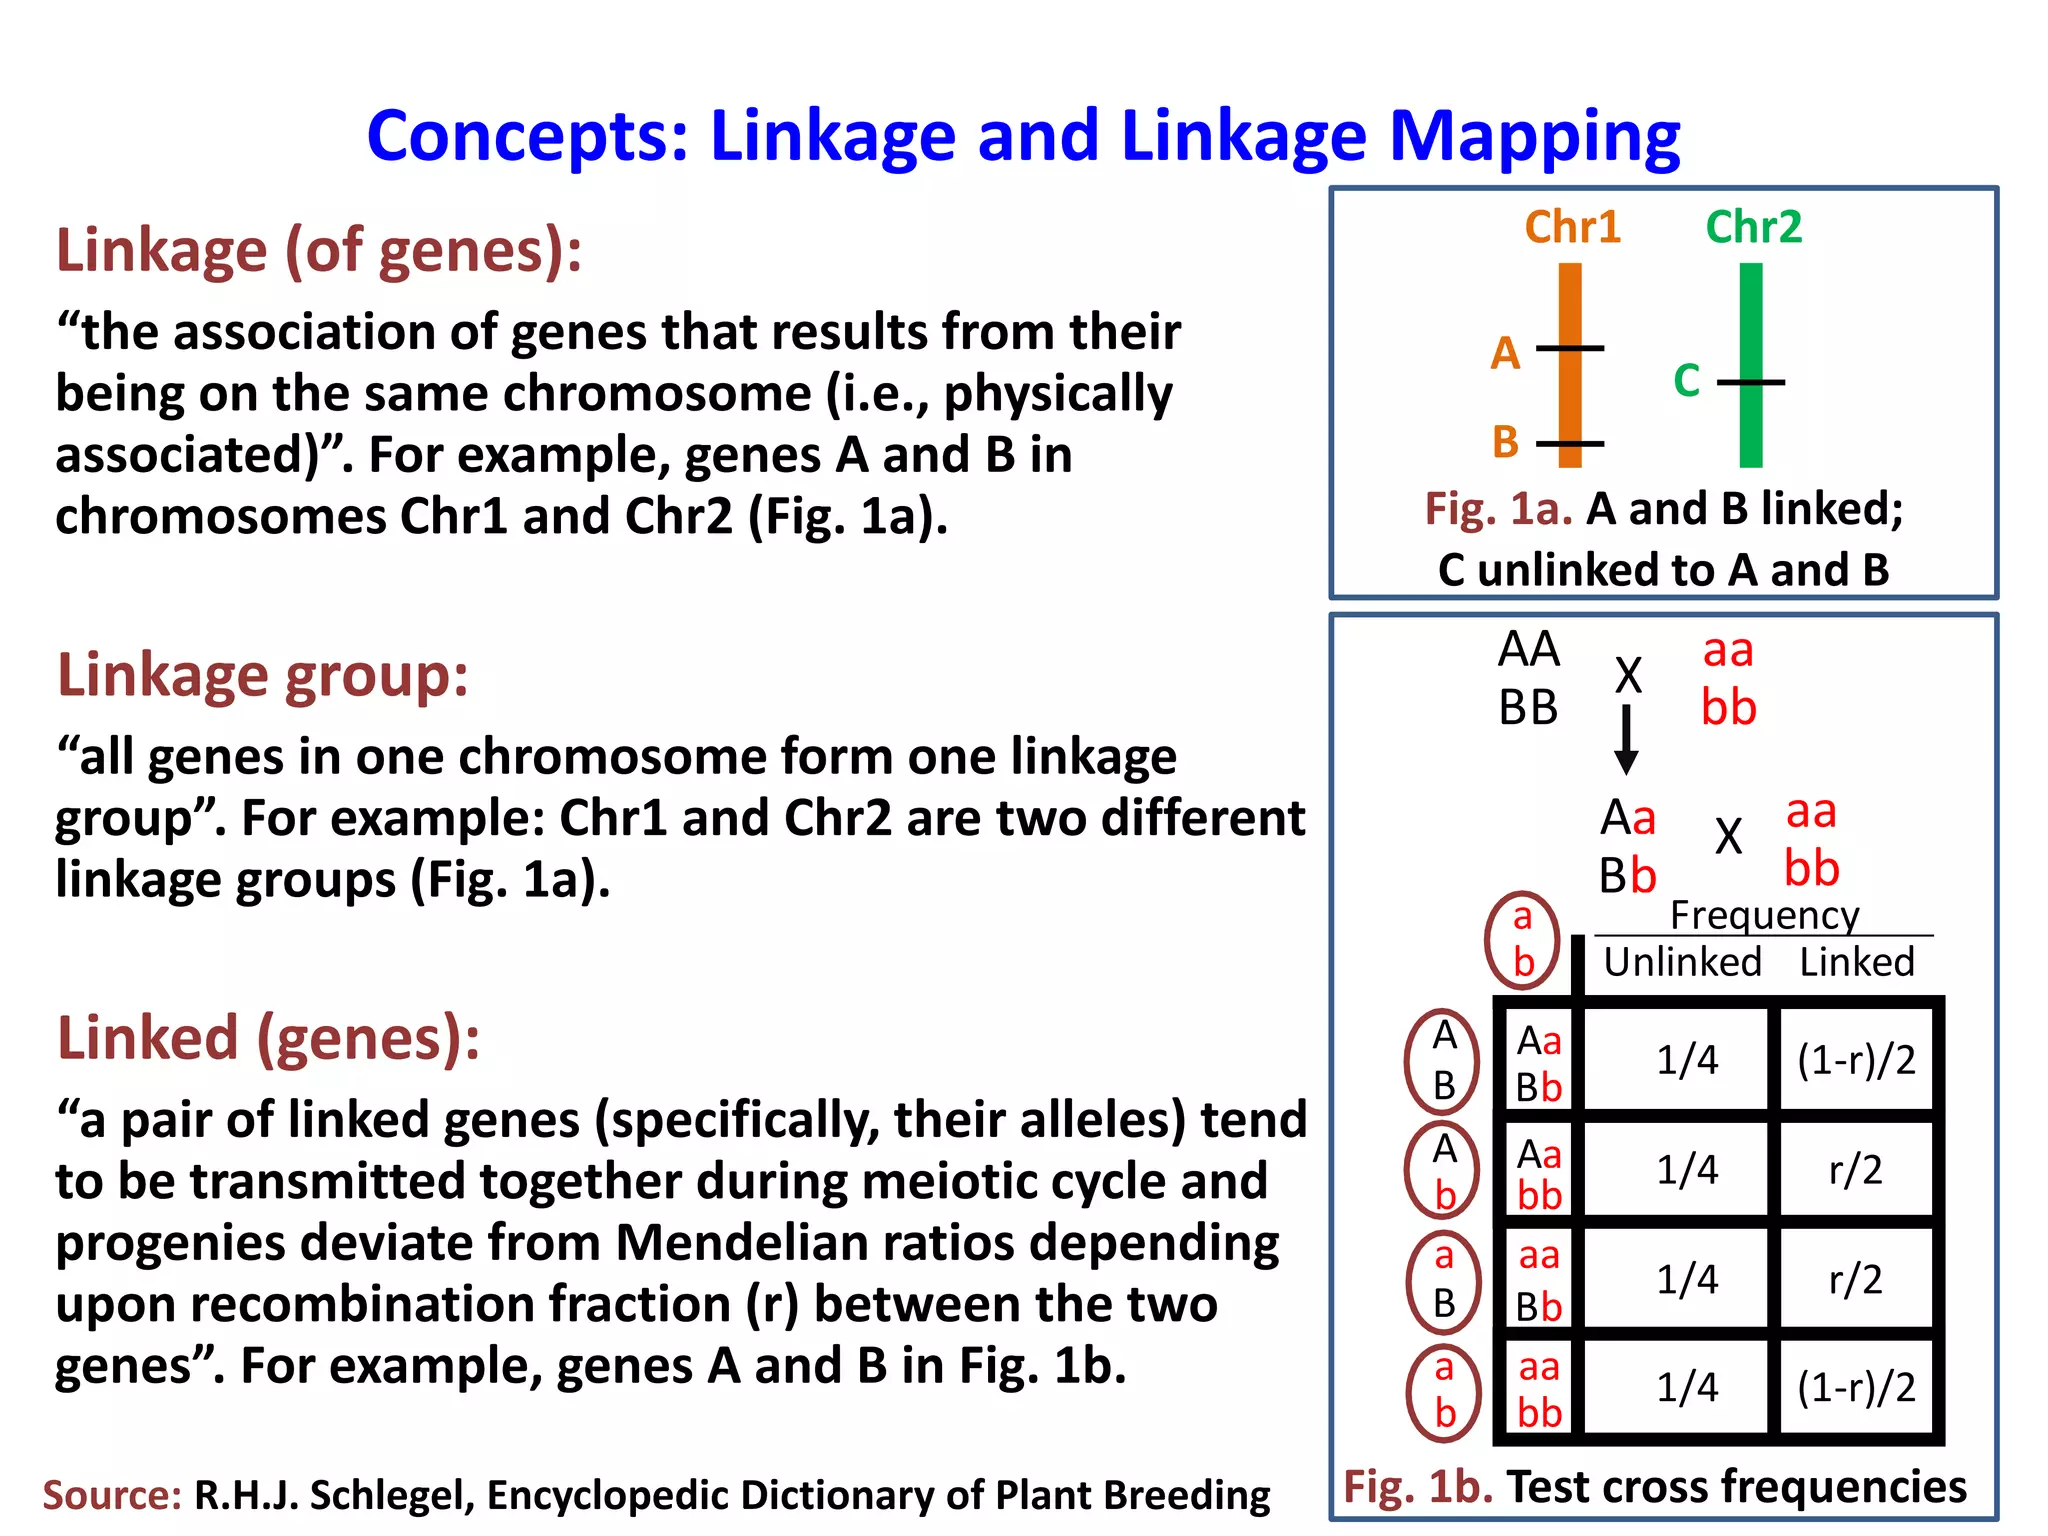
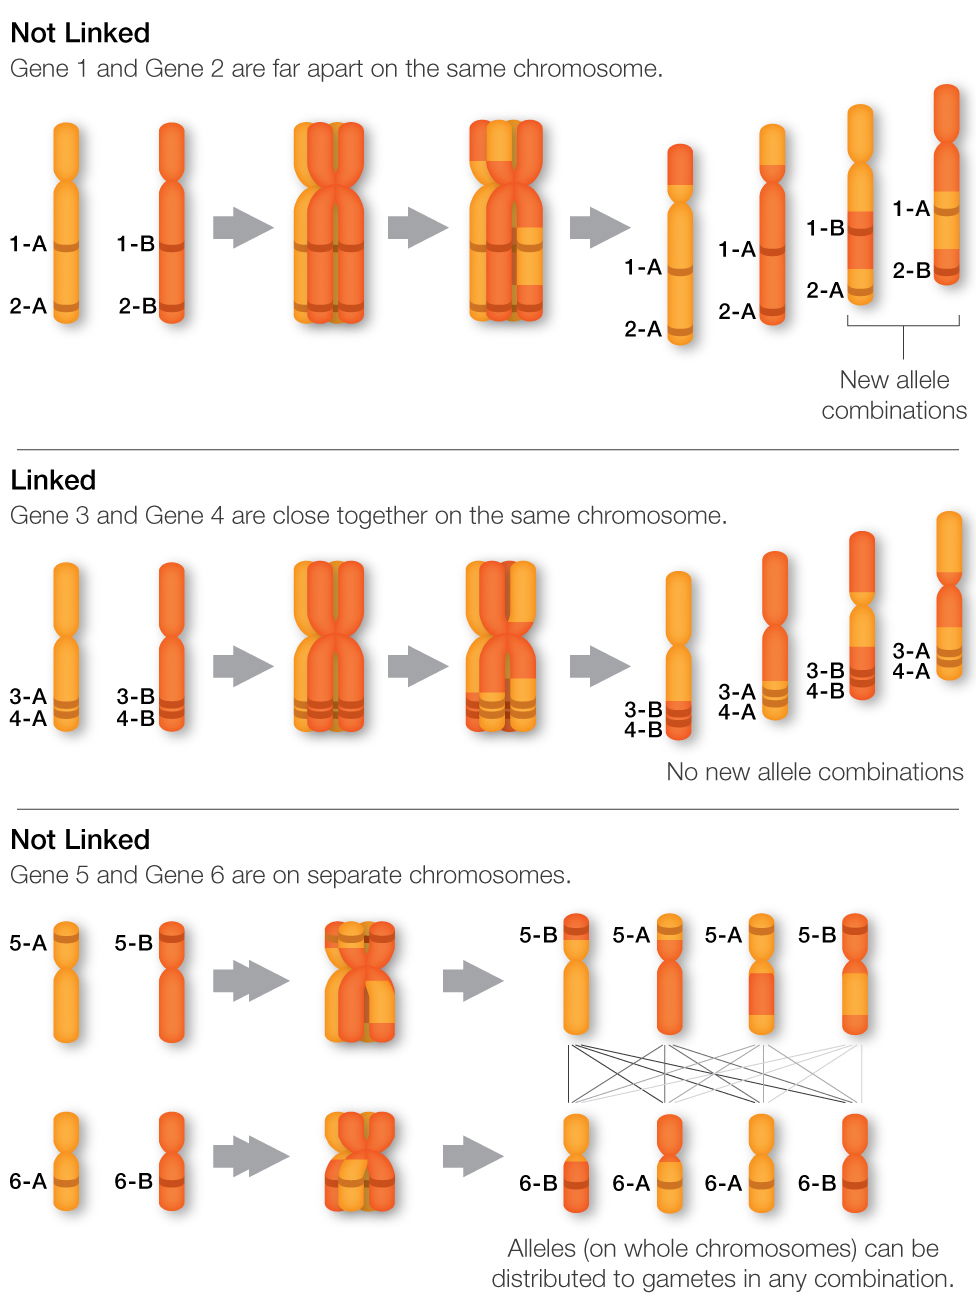
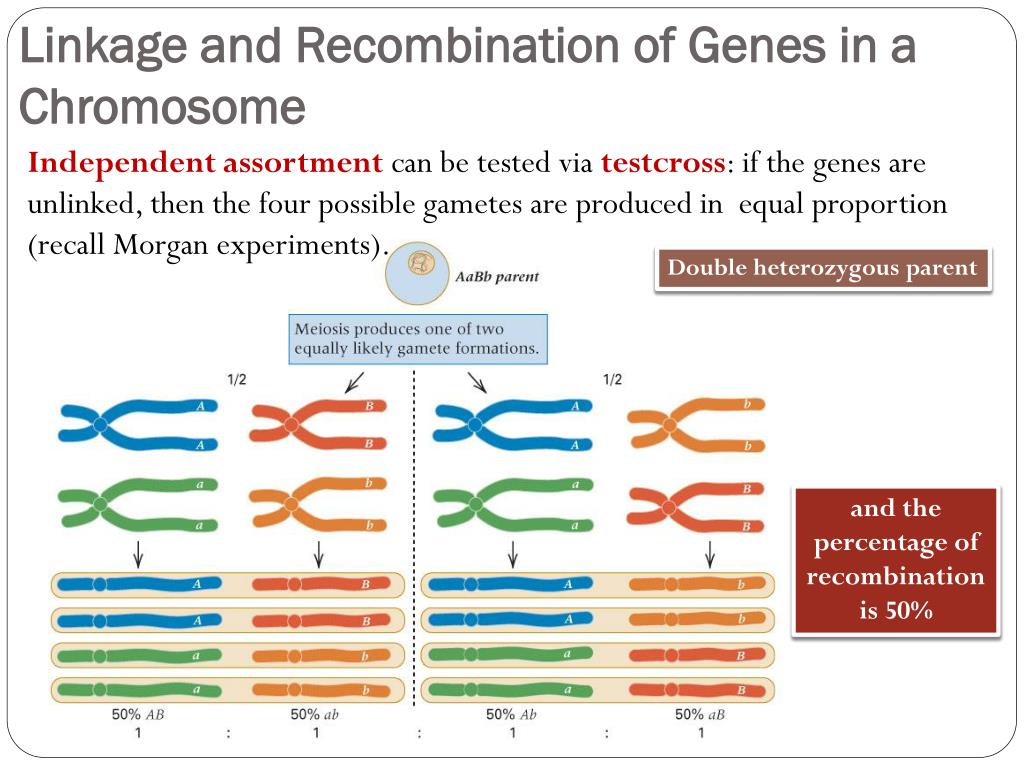
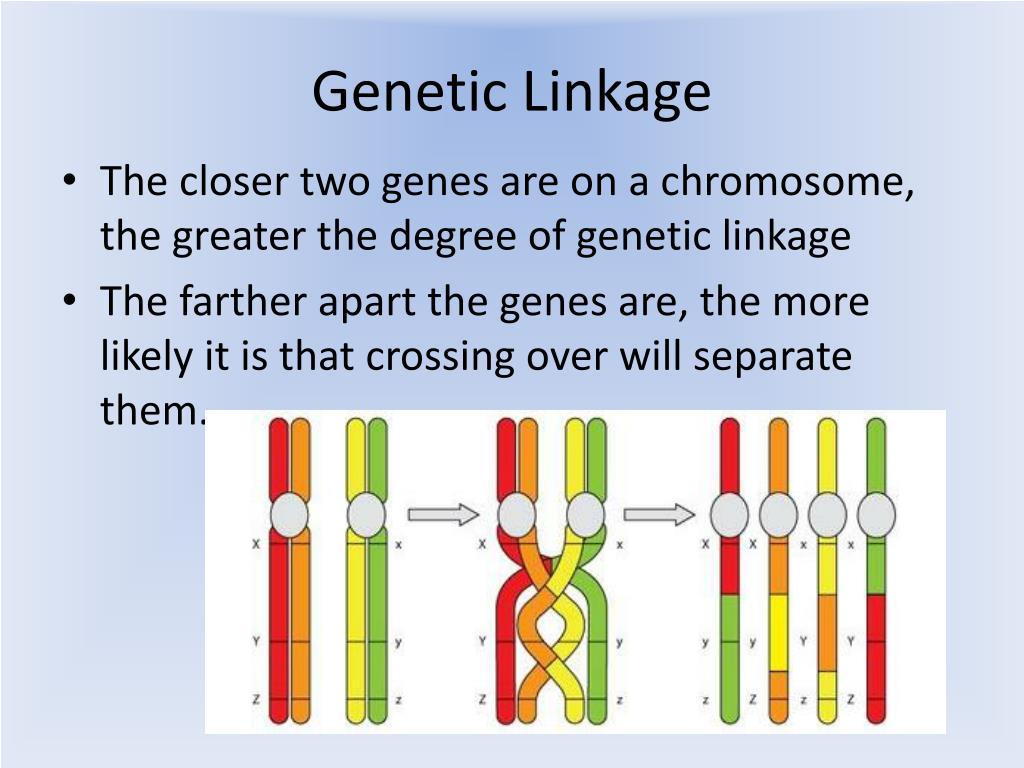
Most people learning genetics hit linkage mapping and immediately get confused about why genes that are close together don't follow Mendel's 9:3:3:1 ratios. The answer is straightforward, but the practical application is where things get messy. I spent a few years working with Drosophila cross data before I stopped treating recombination frequencies like they were exact measurements. Genes located on the same chromosome tend to be inherited together. This isn't a philosophy statement, it's just geometry. During meiosis, homologous chromosomes pair up and can exchange segments through crossing over. The physical distance between two loci determines how often a crossover event separates them. A single crossover between gene A and gene B produces recombinant gametes, while no crossover produces parental-type gametes. The recombination frequency is calculated as the number of recombinant offspring divided by the total offspring, multiplied by 100 to get map units, also called centimorgans. One percent recombination equals one centimorgan. That's the textbook version. In practice, you're working with finite sample sizes and real biological noise.
Here's what the textbooks don't emphasize enough: recombination frequency is not additive over long distances. Two genes 40 cM apart will show less than 40% recombination because double crossovers between them restore the parental configuration. You can't just add up smaller intervals and expect the total to match. This is why mapping functions exist.
How to Actually Build a Linkage Map From Scratch
Start with a controlled cross where you can score phenotypes clearly. A testcross is the standard approach. Cross a heterozygous individual with a homozygous recessive one. The phenotype of each offspring directly reveals the gamete produced by the heterozygous parent, so you don't need anything fancy to read the data. I worked with a set of five markers on chromosome arm 2L in Drosophila melanogaster. The initial plan was to do pairwise recombination frequencies and arrange them linearly. What actually happened is that three of the markers showed apparent recombination frequencies above 50%, which should theoretically mean they're unlinked. They were on the same chromosome. What I was seeing was undetected double crossovers inflating the distance estimates. The workaround was to use a three-point testcross instead of relying on pairwise analysis alone. A three-point cross lets you identify double crossover classes directly because they have the lowest frequency. Once you know which class those are, you can correct the distance estimates by adding twice the double crossover frequency to each relevant interval. This compressed the apparent distances substantially and gave you a map that actually made physical sense.
Get the Full Details
After scoring, here's the step-by-step process I use now: First, determine the parental types by finding the two most frequent phenotypic classes. These represent the non-recombinant gametes. Second, identify the double crossover classes, which will be the two least frequent. Third, use the double crossover phenotype to figure out which gene is in the middle. The middle gene will be the one that's "swapped" relative to the parental configuration in the double crossover class. Fourth, calculate single crossover frequencies for each pair of adjacent genes by counting recombinants in the appropriate intervals. Fifth, convert those frequencies to centimorgans and draw the linear map.
Linked Genes And Linkage Mapping: The Details That Break Your Map
Several things can make your linkage map wrong without any obvious warning sign. I'll cover the ones that actually matter in a real lab setting. Sample size is the biggest factor. With fewer than 200 offspring, your recombination frequency estimates can swing by several percentage points purely from sampling error. A true 15 cM distance might look like 10 cM or 20 cM depending on how the dice roll. I started requiring minimums of 400 scored progeny for any map I intended to publish or use for further crosses. Marker density creates another problem. If your markers are too far apart, you miss the double crossovers entirely and underestimate distances. If they're too close, you can't distinguish them from zero recombination. A typical sweet spot for Drosophila work is markers roughly 10 to 20 cM apart. You can refine later by adding intermediate markers once you have a scaffold map.
Sex-specific recombination is something people forget. In Drosophila, males show zero recombination. If you accidentally use male heterozygotes in your cross, you'll get exactly 0% recombination for every pair and conclude everything is linked when you've actually learned nothing. Always confirm which sex undergoes recombination in your organism before setting up the cross. In humans and mice, females have higher recombination rates than males, which means your map will differ depending on which parent's meioses you're analyzing.
When Linkage Mapping Fails Completely
There are scenarios where this method is simply not viable, and it's worth knowing them early rather than discovering them after months of crosses. Lethal alleles are the main killer. If a particular combination of alleles causes embryonic death, you'll never see those genotypes in your offspring. The missing classes distort your recombination frequencies in unpredictable ways. I once spent three weeks trying to resolve an inconsistent map order before realizing a linked recessive lethal was culling specific recombinant classes. The workaround was to maintain the lethal allele in a balanced stock using a chromosomal inversion and score only the viable classes with the expected distortion factored in. Structural variants like inversions suppress recombination. If an inversion is present in one parent, crossover products within the inverted region produce unbalanced gametes that are often inviable. This makes the region appear completely linked even though the genes are physically far apart. Karyotyping or PCR genotyping for known structural variants before starting a mapping project saves you from chasing phantom linkages.
For organisms where controlled crosses aren't possible, linkage mapping still works but requires population-level approaches instead of experimental crosses. You use existing genetic diversity in a population and look for linkage disequilibrium. The principle is identical, but the resolution is worse and you need hundreds or thousands of samples. Human genetics does this constantly. Population-scale LD decay patterns tell you about historical recombination rates across generations rather than clean meiotic events from a single cross.
Software and Tools I Actually Use
You don't need to hand-calculate everything anymore. I use JoinMap for constructing linkage groups from multi-parent populations and R/qtl for mapping quantitative trait loci in experimental crosses. Both handle the double crossover correction automatically and give you confidence intervals on your map positions. For quick two-point checks, I just write a short R script that takes a phenotype table and spits out recombination fractions with Fisher's exact test p-values. Mapping software does one thing better than manual calculation and that's handling missing data. Real experiments always have missing genotypes. Some offspring fail to score cleanly, some DNA samples degrade, some classes are ambiguous. Software lets you specify likelihood methods that integrate over missing data rather than dropping individuals and biasing your sample. Dropping missing data is the fastest way to introduce systematic error into your map.
Practical Workflow for a New Mapping Project
Here's what I recommend if you're starting a linkage mapping project and want to avoid the mistakes I made. Plan your crosses so you can score at least 500 progeny. Choose markers spaced roughly 10 to 20 cM apart based on previous maps or physical sequence data if available. Use a three-point or multipoint design rather than relying on pairwise analysis. Verify that your markers don't fall inside known inversions or near lethal loci. Run the data through JoinMap or R/qtl and inspect the likelihood curves manually rather than trusting the default output. Check for sex-specific differences if your organism has heterogametic sex determination. Repeat the mapping with an independent population if possible, because first-pass maps always have minor errors that become obvious only after a second round. The whole process from cross setup to a published map usually takes between eight and sixteen weeks depending on the organism's generation time. Drosophila moves fast, so I've seen complete two-point maps generated in under three weeks with dense marker sets. Plants like Arabidopsis take longer because of generation time, but the biology is cleaner. You get uniform selfing and easily generated F2 populations. Linkage mapping is still the foundation of genetic analysis even though whole-genome sequencing exists. Sequencing tells you the physical sequence. Linkage mapping tells you the functional genetic architecture. The two approaches answer different questions. Knowing how to build and interpret a linkage map remains a basic requirement for anyone working in genetics, whether you're studying inheritance patterns in fruit flies or working on crop improvement programs. The theory is simple. The execution is where experience matters.