The Lowry Theory Of Acid And Base
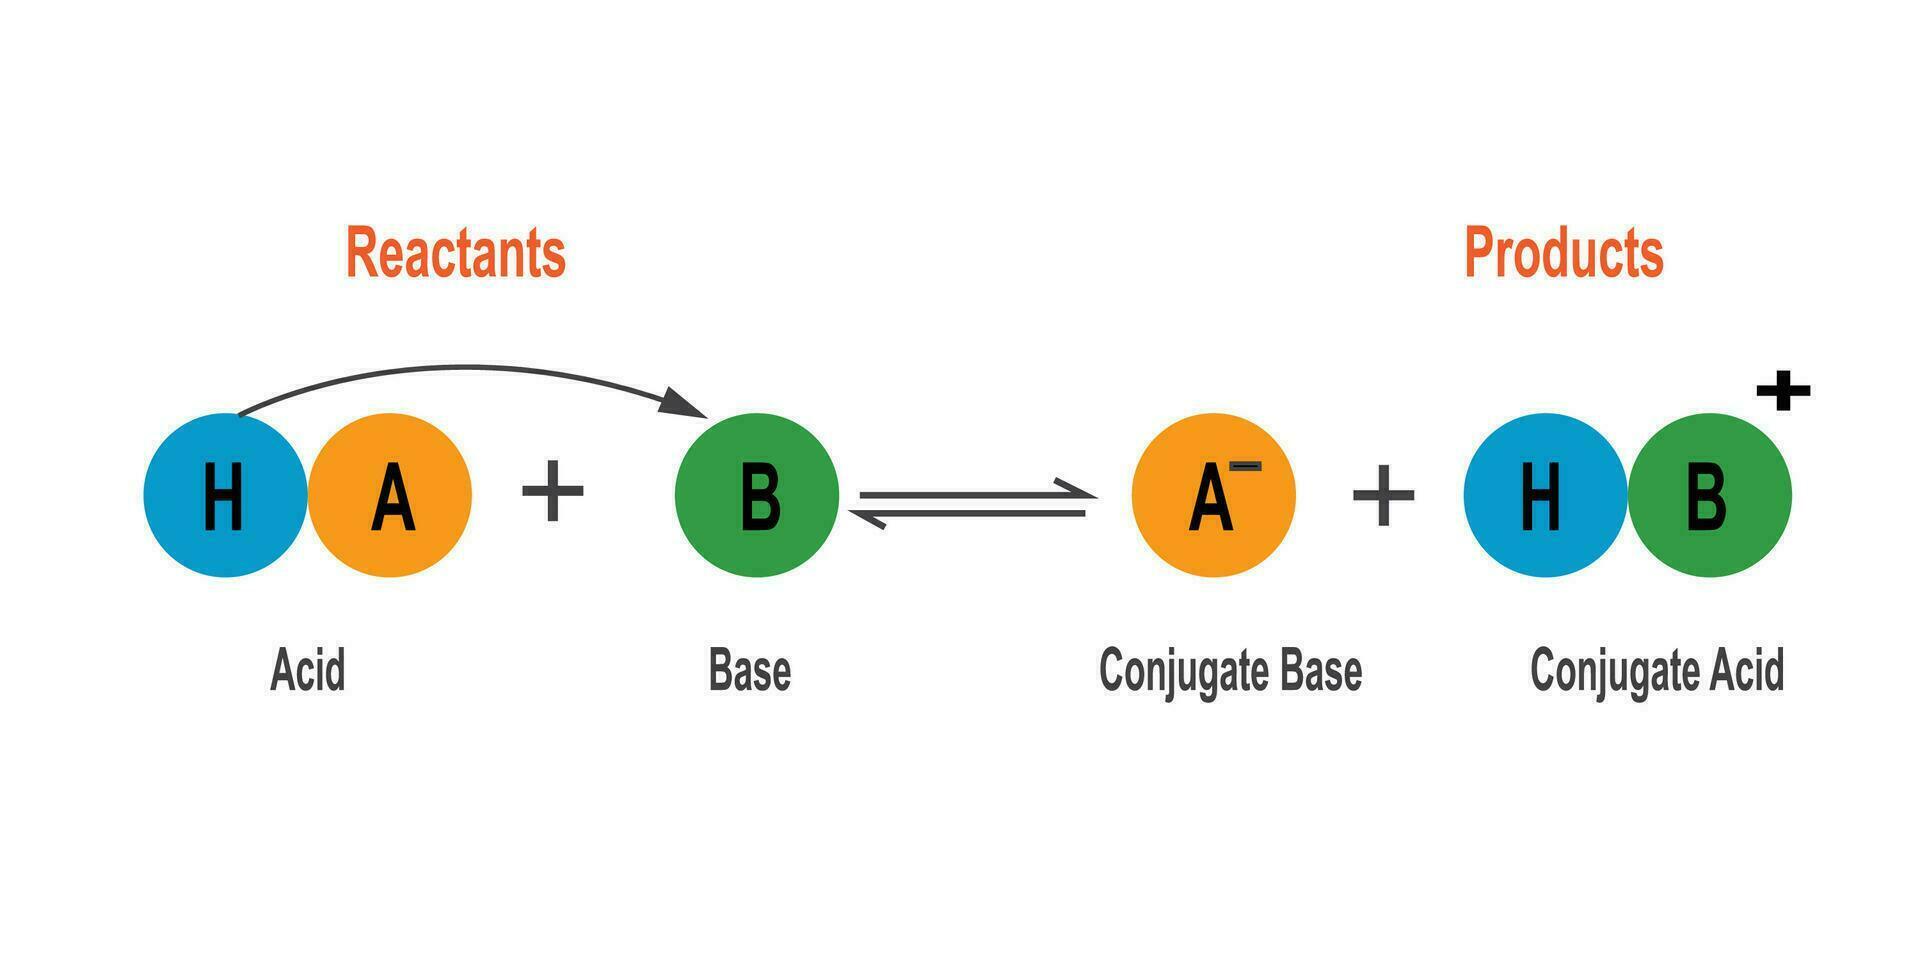
The Bronsted-Lowry definition of acids and bases has been the standard for teaching since the 1920s, and honestly, it still holds up fine for the vast majority of practical work you'll encounter. The core idea is simple: an acid donates a proton, and a base accepts a proton. That's the entire framework. Everything else follows from that. What makes this definition useful compared to the Arrhenius version is that it doesn't require water. Arrhenius only works when you're already in aqueous solution, which means you can't explain why ammonia acts as a base in benzene or why liquid ammonia is a common solvent for inorganic reactions. Bronsted-Lowry handles all of that without modification because it's purely about proton transfer, not about ionization in water.
Lowry Theory Of Acid And Base
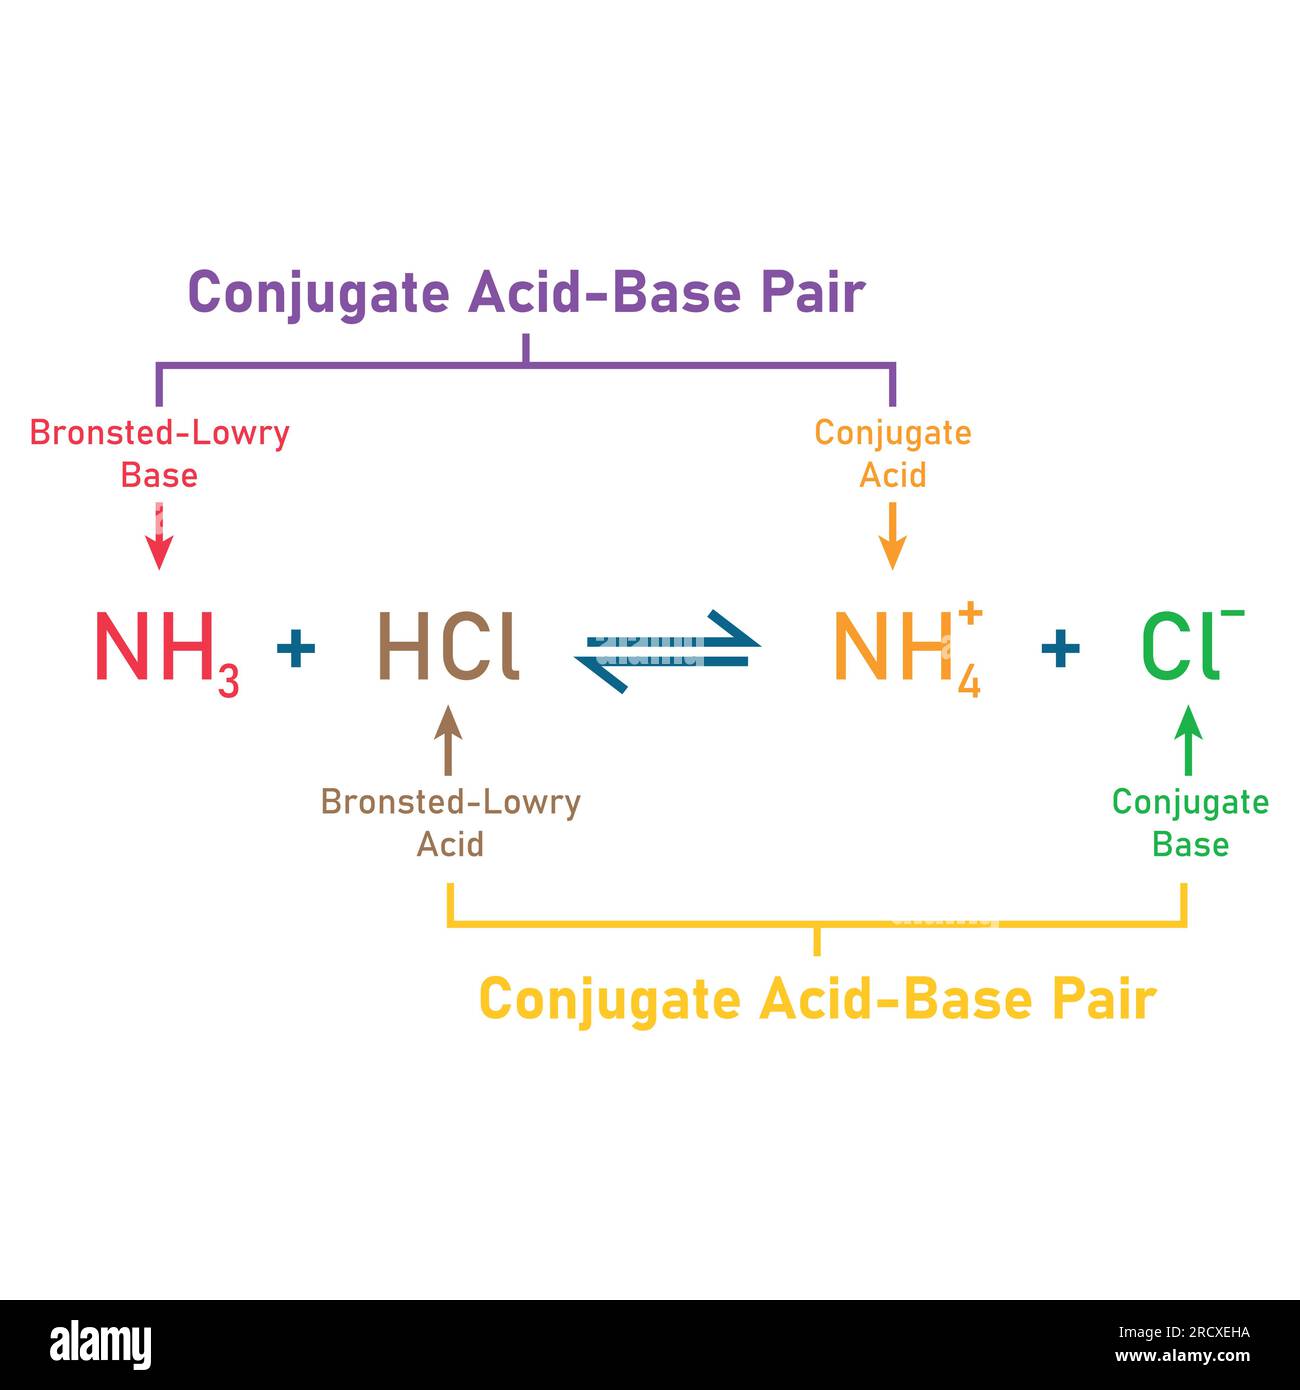
Here's how you actually apply it when you're sitting at a bench and need to figure out what's going on in a reaction mixture. Look at every species involved and ask which one is losing a proton and which one is gaining one. That's it. The species that loses the proton becomes its conjugate base. The species that gains the proton becomes its conjugate acid. These pairs are linked, and tracking them correctly is where most people mess up. Take the reaction between acetic acid and ammonia in water. Acetic acid donates a proton to ammonia. The acetic acid turns into acetate ion. The ammonia turns into ammonium ion. You've got two conjugate pairs: CH3COOH/CH3COO and NH3/NH4+. Write them out explicitly before you move forward. It saves you headaches later when you're dealing with equilibria calculations. One thing beginners consistently get wrong is assuming the stronger acid always means a faster reaction. Acid strength is about equilibrium position, not reaction rate. A very strong acid can participate in a sluggish reaction if the mechanism has a high activation barrier. Don't conflate pKa with kinetics. They're related but they're not the same thing.
How to Use It in Practice
When you're working through problems involving polyprotic acids, the stepwise Ka values matter more than people realize. For something like phosphoric acid, Ka1 is about 7.5 times 10 to the negative 3, Ka2 is 6.2 times 10 to the negative 8, and Ka3 is 4.8 times 10 to the negative 13. The gaps between these values are so large that in most practical calculations you can safely ignore the later dissociation steps. Treat H3PO4 as a monoprotic acid for initial pH estimation, then refine if your precision requirement demands it. Trying to solve the full cubic equation for a triprotic system is almost never necessary outside of analytical chemistry coursework. I ran into a specific problem last year while trying to back-titrate an unknown amine sample using the Bronsted-Lowry framework. The compound was a secondary amine dissolved in an ethanol-water mixture, and I kept getting inconsistent equivalence points. The issue turned out to be that ethanol is a weak acid itself, and at high concentrations it was competing for the base. The titration curve showed two inflection regions instead of one clean endpoint. I switched to using a higher water-to-ethanol ratio and added a smaller volume of titrant at each increment near the suspected endpoint, which brought the inflection point into focus. It took three extra hours of work but gave reproducible results within 2 percent. Another nuance that doesn't get enough attention is the concept of leveling solvents. In water, any acid stronger than hydronium gets completely converted to H3O+. You can't distinguish between the strength of perchloric acid, hydrochloric acid, or nitric acid in aqueous solution because they all level to the same effective strength. If you need to rank their actual acidities, you have to use a less basic solvent like acetic anhydride or glacial acetic acid. The same applies to bases. Any base stronger than hydroxide levels to OH minus in water.
Get the Full Details

When This Framework Breaks Down
The Bronsted-Lowry theory has real limitations, and you should know them before you rely on it blindly. It cannot explain acid-base behavior in systems where no proton is involved at all. Lewis acid-base theory covers those cases, like the reaction between boron trifluoride and ammonia, or the interaction of metal cations with ligands. If your reaction doesn't involve hydrogen transfer, the Lowry theory simply doesn't apply, and you're better off switching to the Lewis definition immediately rather than forcing a proton-based explanation that won't work. Another practical limitation is that proton transfer equilibria in non-aqueous solvents are much harder to quantify. pKa values shift significantly depending on the solvent, and reference tables for organic solvents are sparse compared to water. If you're working in acetonitrile or DMSO, you can't reliably use aqueous pKa values without correction factors. DMSO in particular is notorious for making apparently weak acids seem dramatically stronger than they are in water. Phenol, for example, has a pKa of about 10 in water but drops to around 18 in DMSO on some scales, which seems contradictory until you account for solvent stabilization effects. Autoprotolysis of the solvent also complicates things. Water autoionizes to give a maximum pH range of about 0 to 14. Other solvents have different autoprotolysis constants that define their own usable ranges. In liquid ammonia, the useful range is roughly 0 to 33 on the pNH scale. Trying to carry over aqueous pH intuition into other solvents without converting to the appropriate scale will give you wrong predictions about whether a given acid or base will even exist in solution.
A Quick Reference for Common Conjugate Pairs
Having a mental list of strong conjugate pairs saves time during problem solving. Strong acids like HCl, HBr, HI, HNO3, H2SO4, and HClO4 all have negligible conjugate bases. Their conjugate bases are so weak they won't accept protons under normal conditions. On the basic side, strong bases like hydride (H minus), amide (NH2 minus), and methoxide (CH3O minus) have conjugate acids that are very weak. Understanding which end of the spectrum you're working with helps you predict reaction direction without doing full equilibrium calculations every time. The key takeaway is that the Bronsted-Lowry model is a tool, not a law of nature. It works extremely well for proton-transfer reactions in protic solvents, which covers most of what you'll encounter in undergraduate labs and routine synthetic work. When you step outside that zone, you need to recognize the boundary and switch frameworks rather than trying to stretch the theory beyond its intended use.