What actually happens when you run a sample through the machine
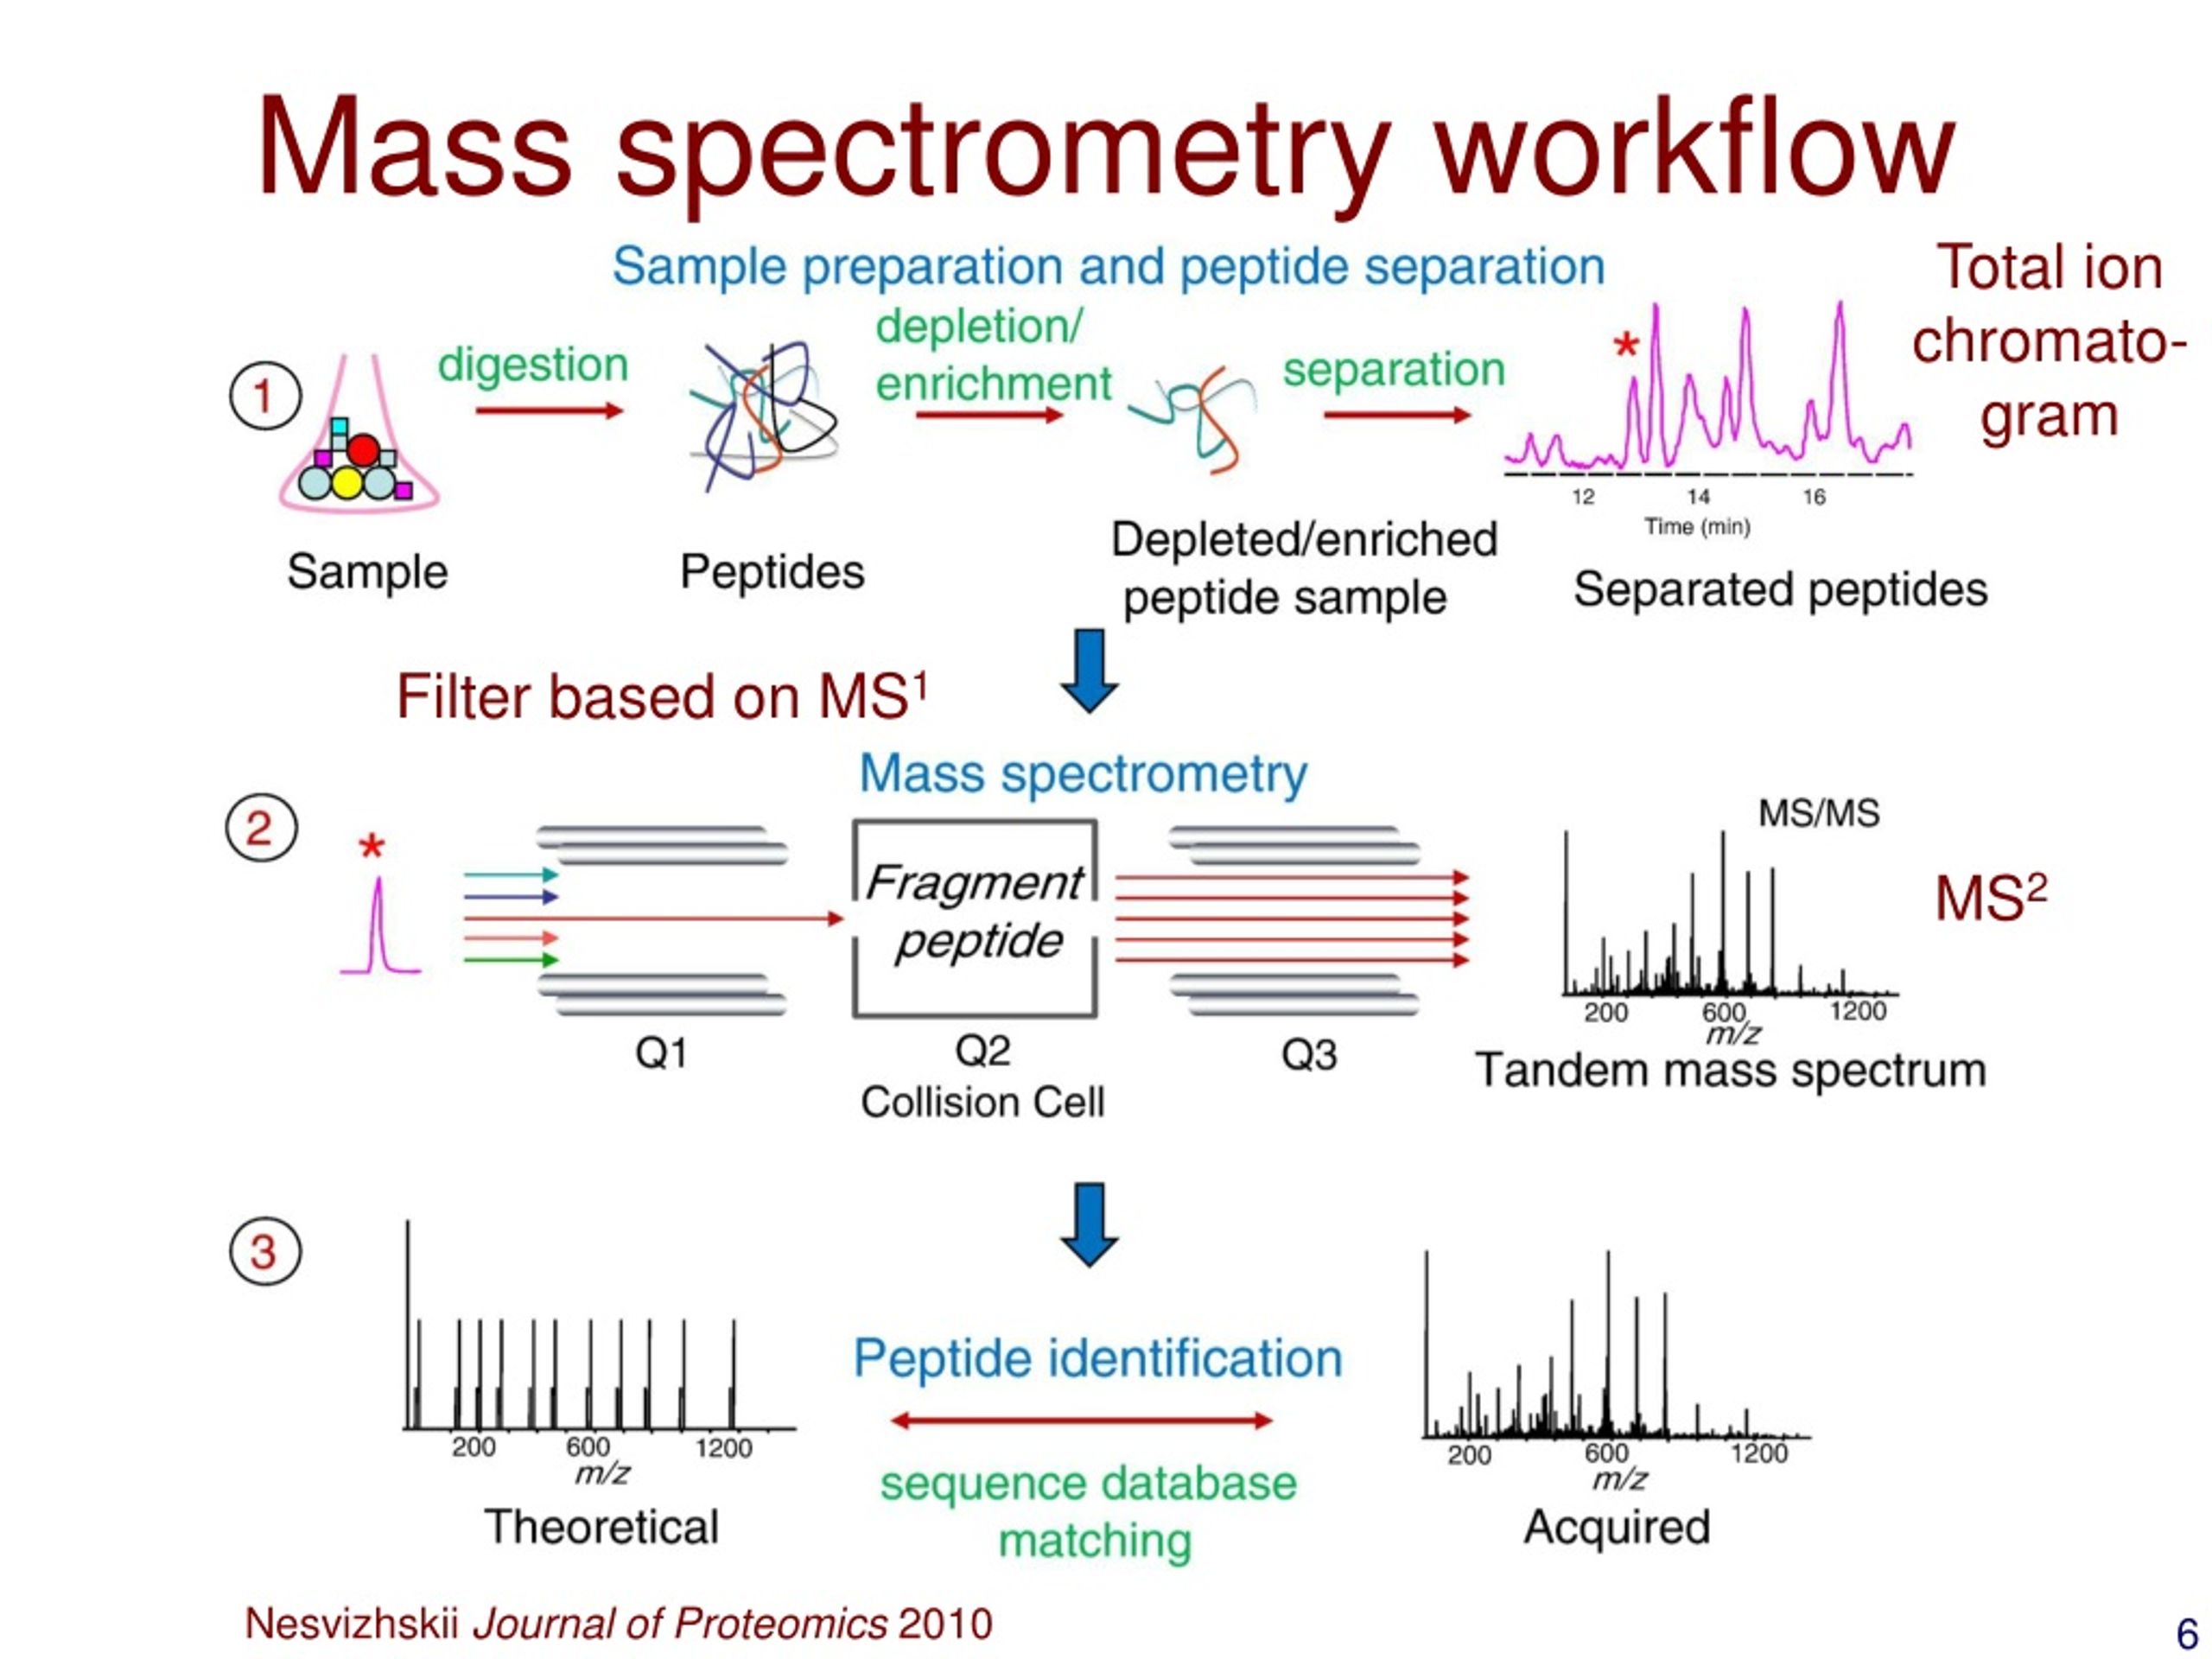
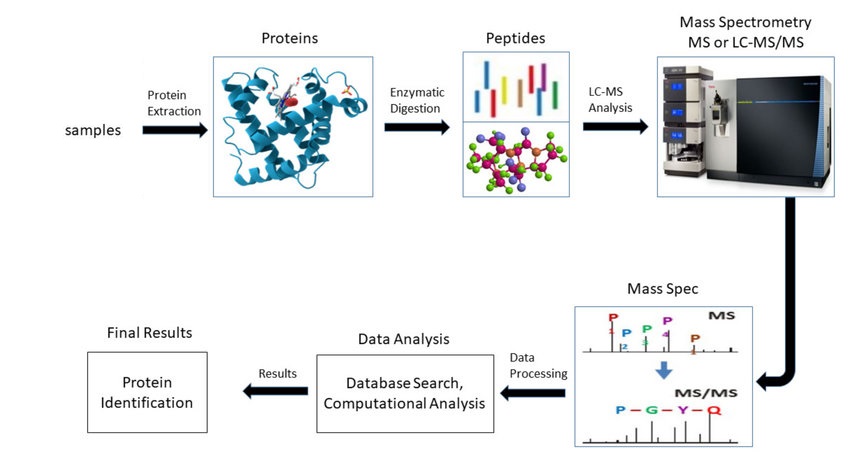
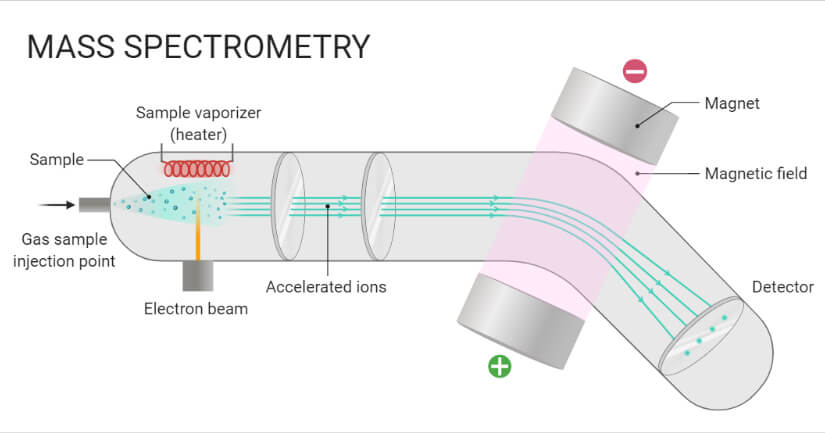
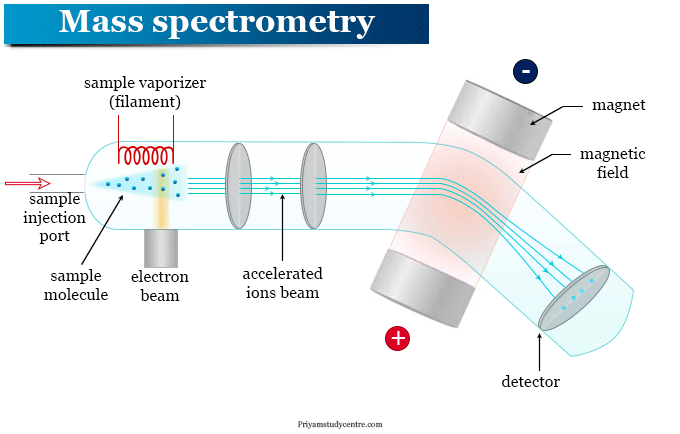
Mass Spectrometry Analysis is a technique that measures the mass-to-charge ratio of ionized molecules. You introduce a sample, it gets vaporized and ionized, the ions travel through a mass analyzer, and a detector records what comes through. The output is a spectrum that tells you the molecular weight, structural fragments, and sometimes the exact composition of whatever you put in front of it. That's the textbook version. The real version involves more frustration. I've spent years working with these instruments in what I'd call industrial quality control settings. We're not doing elegant structural biology here. We're running thousands of samples a week trying to figure out if a batch of synthetic intermediates has the right impurities, or whether a pharmaceutical compound is degrading in storage. The theory is clean. The practice is messy.
Mass Spectrometry Analysis in practice: the ionization question
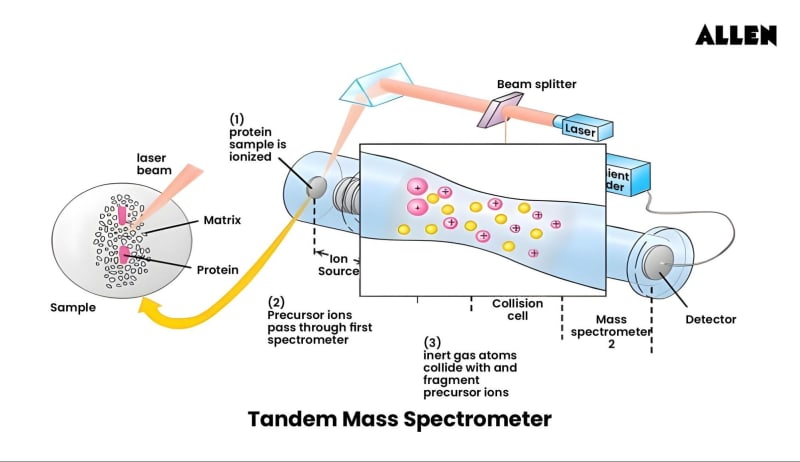
The first decision you make before running anything is how to ionize your sample. This matters more than most beginners realize. Electrospray Ionization (ESI) is the default for polar, thermally labile compounds. It produces mostly singly or doubly charged molecular ions, which is why it pairs so naturally with liquid chromatography. Soft ionization, minimal fragmentation, clean molecular weight data. Good for proteins, peptides, polar metabolites, anything water-soluble. But ESI chokes on nonpolar compounds. I've seen people waste entire days trying to get signal from hydrophobic small molecules in pure ESI mode. Switch to Atmospheric Pressure Chemical Ionization (APCI) and the same compounds roar to life. APCI is harder ionization by nature. It uses a corona discharge needle to create reagent ions that then transfer charge to your analyte. You get more fragment ions, less [M+H]+ clarity, but you actually see something instead of baseline noise. MALDI is the third major player and it lives in a different world. You co-crystallize your sample with a matrix compound on a steel plate, then hit it with a laser pulse. It's the go-to for large biomolecules, polymers, and imaging applications. The downside is that MALDI spectra are notoriously sensitive to matrix crystallization quality. Two people running the same sample on the same instrument can get wildly different results depending on how their crystals formed. I spent three weeks troubleshooting a peptide MALDI problem before I realized the acetone in my matrix solution had absorbed enough water from the air to shift the crystallization kinetics. Freshly prepared matrix solved it immediately.
The analyzer types and why they matter more than the detector
Your mass analyzer determines resolution, mass accuracy, scan speed, and dynamic range. These are not independent variables. You pick what you need and accept what you lose. Quadrupole instruments are the workhorses. Single quad, triple quad, quadrupole time-of-flight. A single quad gives you unit resolution, usually around 0.7 to 1.0 Da peak width at half height. Useful for targeted quantification with selected ion monitoring, but you're not identifying unknowns with high confidence. Triple quads, or tandem MS, are the gold standard for targeted quantitation in complex matrices. The first quadrupole selects a precursor ion, the collision cell fragments it, and the third quadrupole monitors specific product ions. This is Multiple Reaction Monitoring, MRM, and it's what regulatory labs use for bioanalysis. You can detect compounds at parts-per-billion or even parts-per-trillion levels in plasma, urine, tissue homogenates. The method development time is the cost. Each transition needs optimization, and you'll spend a week getting clean MRM transitions for a new analyte if you're doing it right. Time-of-flight analyzers measure the time it takes ions to travel a fixed distance. Faster ions arrive first. High-resolution TOF gives you sub-ppm mass accuracy, usually under 2 ppm with proper calibration. This changes what you can do with unknown identification. You can determine elemental compositions from accurate mass alone in many cases. Orbitrap analyzers work differently. They trap ions in an electrostatic field and measure the frequency of their oscillations. Resolution can reach 140,000 or higher at m/z 200. Mass accuracy routinely hits 0.5 to 1 ppm. If you're doing proteomics, metabolomics, or fine chemical characterization, Orbitrap is where you want to be. The tradeoff is scan speed. Modern Orbitraps are fast, but they still can't match the acquisition speed of a good triple quad for targeted work.
Get the Full Details
Ion traps are worth mentioning even though they've lost market share. They're compact, relatively inexpensive, and excellent for MSn experiments. You can do three or four stages of fragmentation on a single trapped ion, which is useful for structural elucidation. But they have a well-known space-charge limit. Put too many ions in and the resolution and mass accuracy degrade. This is a practical problem, not a theoretical one. I've seen analysts push ion traps past their limits trying to save time between injections, then wonder why their mass accuracy drifted from 1 ppm to 10 ppm over the course of a batch run. Keep your ion populations reasonable and recalibrate between batches.
Sample preparation: where most problems actually start
People obsess over instrument parameters and method optimization, but the majority of bad data comes from poor sample prep. A clean extract runs well on any instrument. A dirty one ruins everything regardless of how expensive your mass spectrometer is. Protein precipitation is the simplest approach and often sufficient. Add three volumes of acetonitrile to your aqueous sample, vortex, centrifuge, inject the supernatant. Works for clean matrices. Fails when you have lipids, phospholipids, or other interferences that co-precipitate poorly and then suppress your ionization. Lipid removal is the single most impactful cleanup step for bioanalytical work. Phospholipids are notorious ionization suppressors in ESI. Even 10 µg of phospholipid per mL of injected sample can reduce your analyte signal by 50 percent or more. Use a phospholipid removal plate if you're doing high-throughput work. One pass through the plate cuts phospholipid content from micrograms to nanograms per injection and improves both sensitivity and instrument cleanliness. Solid-phase extraction remains the most flexible cleanup method. You can tailor the sorbent to your analyte's chemistry. C18 for reverse-phase retention of moderately polar compounds, mixed-mode cation-exchange for basic drugs, HLB for broad-spectrum retention. The trick is choosing the right equilibrium conditions. If your sample is too aqueous, C18 won't retain anything. If it's too organic, polar impurities will co-elute and contaminate your spectrum. Test your retention with a small aliquot before committing the full sample.
Here's a specific problem I dealt with that took far too long to diagnose. We were running a stability-indicating method for a basic pharmaceutical compound using LC-MS/MS. The method worked perfectly on fresh standards. But every environmental sample showed an unexpected peak at the same retention time as our analyte, with identical MRM transitions. It took two weeks of investigation before I realized the issue was column bleed from a degraded C18 column producing a dimethyl siloxane cluster ion that happened to share the same precursor and product masses as our compound's fragment. Same mass transition, different chromatographic behavior, but we were looking at extracted ion chromatograms and assuming co-elution. Switching to a fresh column eliminated the interference. It wasn't a real impurity at all. It was the column talking back.
Data processing and interpretation without fooling yourself
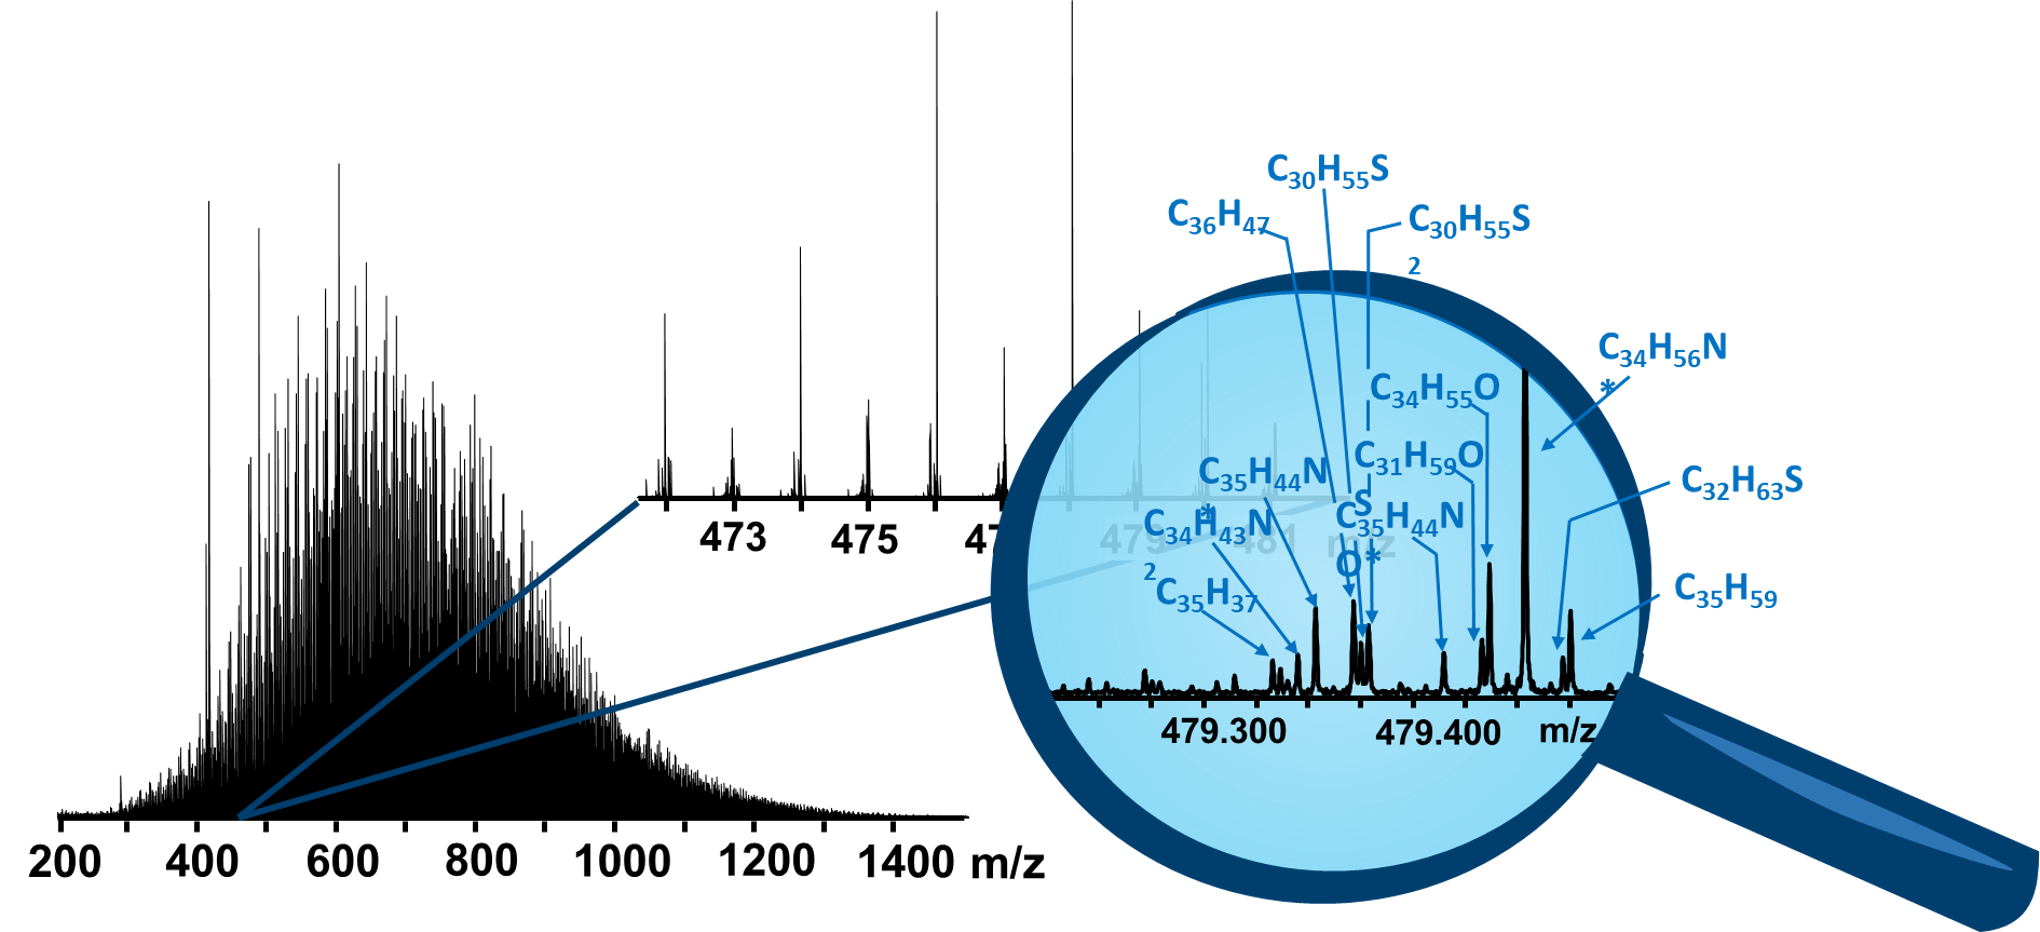
Modern instruments generate enormous amounts of data. Processing it correctly requires discipline. The biggest trap is confirmation bias. You expect to see a certain peak at a certain retention time, so you interpret ambiguous signals as confirmation rather than questioning them. Use internal standards. Isotopically labeled internal standards are ideal because they co-elute with your analyte and experience the same matrix effects. If you're doing quantification without an internal standard, your results are only as reliable as the stability of your instrument response over the entire run. That's rarely good enough for regulated work. Matrix-matched calibration curves help, but they don't correct for ionization suppression that varies between individual samples. An internal standard corrects for both. For qualitative work, accurate mass data alone can identify compounds in many cases. Use the mass accuracy to generate a list of possible elemental compositions, then filter against your knowledge of the sample. A compound synthesized from starting materials A, B, and C shouldn't contain sulfur unless one of those starting materials has sulfur in it. Use your chemistry to constrain the possibilities. High-resolution data from an Orbitrap or TOF instrument makes this feasible. Unit-resolution data from a single quad does not.
Fragmentation patterns are the next layer of information. Collision-induced dissociation, CID, produces structural fragments that can confirm identity beyond what mass alone can do. Compare your experimental spectra to reference libraries when available. NIST and mzCloud have extensive collections. But don't trust library matches blindly. A cosine similarity score of 900 out of 1000 sounds convincing until you realize the reference spectrum was acquired under completely different collision energy conditions. Match scores are not definitive identification. They're supporting evidence.
Common failures and what to do about them
Mass spectrometers are sensitive instruments that require regular maintenance. Here are the issues I encounter most frequently, listed roughly by how much they frustrate people. Signal drift is usually contamination. Ions are sticking to surfaces inside the interface region. Clean the ion source according to the manufacturer's procedure. For ESI sources, this typically means removing the spray shield, capillary, and skimmer cones and soaking them in a appropriate solvent. Acetonitrile with 1 percent formic acid works for most organic residues. For protein deposits, try a weaker detergent solution or specialized cleaning protocol. Reassemble and run a tuning standard. Signal should recover within 15 to 30 minutes of stable operation. Mass accuracy drift points to calibration issues. Lock mass correction helps but doesn't replace periodic calibration. Run a calibration standard at the beginning and end of every batch. If the drift exceeds your acceptance criteria, recalibrate mid-batch. For Orbitrap instruments, check the calibration file integrity. Corrupted calibration files are a real problem and not as rare as you'd think after a power fluctuation or software update.
Contaminant peaks appearing everywhere usually trace back to solvents or consumables. LC-grade solvents are not the same across brands. Some contain impurities that show up as background peaks in your mass spectrum. Test your solvents by running a blank injection on your method before committing to a new lot. Disposable tips and vials can leach plasticizers and siloxanes. Use low-bind, certified-low-background consumables for trace analysis. The cost difference is small compared to the cost of re-running a batch because your vial caps are contaminating your samples. Source contamination from complex matrices is inevitable. The solution is to inject fewer micrograms of total organic material per run. Dilute your samples. Use stronger wash solvents in your LC method. Consider a guard column. These all reduce the rate of contamination accumulation and extend the time between required maintenance cycles.
When Mass Spectrometry Analysis isn't the right tool
It's important to acknowledge the limitations. Mass spectrometry requires your analyte to be ionizable. Some compounds simply don't ionize well in any mode. Highly fluorinated molecules, some perfluoro compounds, and certain polymeric materials produce weak or no signal. You can sometimes force ionization with specialized reagents or alternative ionization techniques, but these require method development and may not be robust. Isomers are another limitation. Standard MS cannot distinguish between structural isomers with the same elemental composition. You need chromatographic separation, ion mobility, or additional structural information from MSn to resolve them. If your sample contains isomeric impurities and you only have a single quad, you're not going to identify them. Pair the MS with a good LC method or move to a higher-resolution instrument with fragmentation capability. Quantification without proper standards is essentially decoration. You can detect a compound, you can get an approximate concentration from a calibration curve, but without a pure standard of the same compound, your numbers are unreliable. This is especially relevant for unknown degradation products or impurities. You can characterize them structurally, but you cannot quantify them accurately without isolating the compound and preparing a calibration standard. This is a constraint that people working in regulatory environments encounter constantly.
NMR spectroscopy complements MS well in these situations. Where MS struggles with isomers and absolute structural confirmation, NMR excels. Where MS excels at sensitivity and throughput, NMR requires more sample and more time. Use both when the stakes are high. Rely on MS alone when you need speed and your compounds are well-characterized. The instrument itself has practical limits. Detection limits are excellent for most applications, typically low nanogram or sub-nanogram per injection for well-ionizing compounds. But if your compound is present at femtogram levels in a complex matrix, you'll need preconcentration or a more sensitive instrument configuration. Some specialized applications push into the attomole range with microflow or nanoflow ESI coupled to Orbitrap instruments, but these require significant optimization and are not routine.
Getting started without making expensive mistakes
If you're new to this, start with what your instrument can do reliably before pushing it to the edge. Learn the maintenance procedures. Learn how to recognize when your data is degrading. Learn the difference between a real analytical problem and an instrument problem. Most instrument issues have signatures. Contamination produces increasing background. Calibration drift shows up as systematic mass error across all peaks. Source deterioration causes gradual sensitivity loss. Learning to read these signs saves more time than any method optimization ever will. Document everything. Instrument parameters, maintenance logs, calibration records, chromatographic conditions, mobile phase preparations, sample preparation details. When something goes wrong six months from now, you'll be glad you wrote it down. I still pull old lab notebooks to trace problems that resurface in modified forms. The specific numbers don't always transfer, but the context does. Don't skip method validation if you're working in a regulated environment. Specificity, linearity, accuracy, precision, limit of detection, limit of quantification, stability, matrix effects. These are not optional checkboxes. They're the difference between data that stands up to scrutiny and data that gets rejected during an audit. The validation process takes time, but skipping it costs more in the long run.
Stay current with the literature, but apply it critically. New ionization techniques, new column chemistries, new data processing algorithms appear regularly. Some are genuinely useful. Many are incremental improvements marketed aggressively. Test everything against your own samples before adopting it into your routine. Your method works for your samples under your conditions. Universal recommendations from papers don't always translate. The field moves fast. Instruments get faster, more sensitive, more automated every few years. But the fundamentals don't change. Good samples, proper ionization, appropriate analyzer, careful data interpretation. Master those and the rest is details.