Setting Up a Material Balance Before You Open Any Spreadsheet
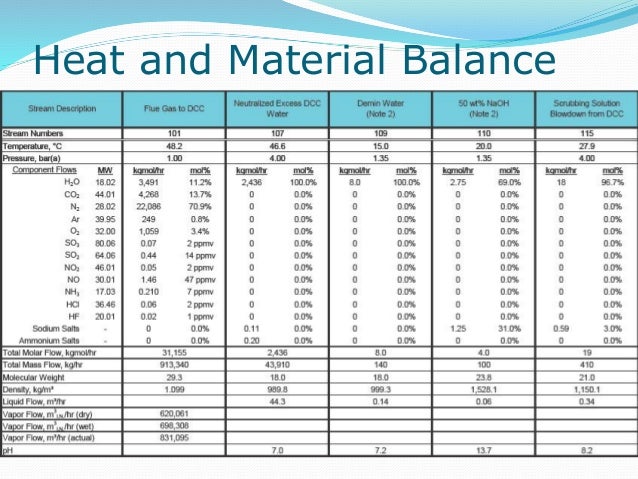
Most people skip straight to writing equations and wonder why their numbers don't close. The actual first step is figuring out what you're dealing with. You need to draw a process boundary and decide exactly what's inside it and what crosses the line. Is your boundary around the whole plant? A single reactor? A flash drum and its heat exchanger? Getting this wrong means you'll spend three hours rearranging equations that were never set up correctly in the first place. I started with a simple steady-state distillation column last year. The specification sheet said 99.5% purity on the distillate and gave me a reflux ratio of 1.8. I tried solving it with total mass balance and component balances on each tray from the top down. By tray four, my numbers were drifting so far off that the column wasn't even physically possible at that reflux ratio. The problem wasn't my math. It was that I hadn't accounted for the non-condensable gases being swept through the top, which I only realized when I looked at the actual P&ID and saw a small vent line I had ignored. Those non-condensables were about 2% of the feed flow but they accumulated at the condenser and changed the vapor-liquid equilibrium enough to throw everything off.Material Balance In Chemical Engineering
The core equation is always the same, no matter how complicated the process gets. Mass in equals mass out plus accumulation. For steady-state operations, which is most textbook problems and the majority of continuous plant work, accumulation is zero. That simplifies it to in equals out. For unsteady state, you add the accumulation term and suddenly you're working with differential equations instead of algebra. General form: Input + Generation = Output + Consumption + Accumulation
In most chemical engineering material balance problems, generation and consumption are zero unless you're doing reactive systems. Then you add mole balances on each species and use stoichiometry. That's where it gets messy fast. The degrees of freedom analysis is where people get tripped up. You count unknowns, you count independent equations, and if they don't match you can't solve it. Unknowns are your stream variables, temperature, pressure, flow rates, compositions. Equations come from balances, equilibrium relationships, design specs, and physical constraints. If your degrees of freedom is positive, the problem is underspecified. Negative means over-specified or redundant. Zero is where you want to be. I've seen engineers try to force solutions on problems with positive degrees of freedom by making arbitrary assumptions that weren't in the problem statement. That's not engineering, that's guesswork.
Reactive Systems and the Real Pitfalls
When reactions enter the picture, you stop balancing mass on individual molecules and start balancing on atomic species or using the extent of reaction method. The atomic species method is more robust because you don't need to know the reaction mechanism. Carbon atoms in equal carbon atoms out. Hydrogen atoms in equal hydrogen atoms out. It doesn't matter if the reaction goes through ten intermediate steps. The atoms don't care. The extent of reaction approach is cleaner when you know the stoichiometry. You define one variable, xi, for each independent reaction and relate every species flow rate back to it. The trick is making sure your reactions are truly independent. If you write three reactions where one is a linear combination of the other two, you've added a dependent equation and your degrees of freedom count is wrong. I once spent two days debugging a syngas balance because someone had written water-gas shift and methanation as independent reactions when they were actually dependent on the hydrogen-oxygen-carbon atom balances. Recycle streams change everything about how you approach a material balance. You can't just solve from inlet to outlet in a straight line anymore. You have to iterate. The standard approach is to tear the loop at a convenient point, guess the recycle flow, solve the rest of the system, compare the calculated recycle to your guess, and update. The convergence speed depends entirely on how you choose the tear stream and what relaxation factor you use. A relaxation factor of 0.5 means you take half the difference between your guess and the new value. Too aggressive and you oscillate. Too conservative and you'll be iterating for hours on a problem that should converge in ten cycles.
Get the Full Details

Unsteady-State Balances
Sometimes you need to model a batch process or a startup transient. The accumulation term becomes important and you're solving differential equations. A simple tank with one inlet and one outlet has a straightforward balance. dV/dt = Qin - Qout. If the densities are constant, volume balance works. If not, you need mass balance and then convert using density. The tricky part with unsteady state is the initial conditions. You need the state of the system at t equals zero, and in practice that's often something you estimate rather than measure. I did a batch reactor transient analysis where the initial concentration was off by 4% because the operator had mixed the charge for only thirty seconds instead of the specified five minutes. The model predicted a 12% higher conversion than what actually happened. The discrepancy wasn't in the kinetics, it was in that initial condition.
Software Tools and What They Get Wrong
Aspen Plus, HYSYS, Chemcad, even Excel with solver. Pick whatever you use and learn its failure modes. These tools will give you an answer quickly, usually in under a minute for a well-posed problem. They will also give you an answer for ill-posed problems and make it look convincing. Always check convergence diagnostics, not just the final numbers. A solver that reports it converged but has a large mass imbalance is worse than useless, it's misleading. Flash calculations in particular are notorious for converging to the wrong root when you have near-critical conditions. I had a problem where the solver kept finding a single-phase solution when the system was clearly in the two-phase region. The issue was that my initial temperature guess was too far from the actual bubble point. Dropping the initial guess by fifteen degrees and tightening the tolerance from one percent to zero point one percent fixed it. The total run time went from a failed convergence to about forty seconds.
Common Mistakes That Waste Time
Forgetting that molecular weights change when you switch between mass and molar units. This is the single most common error I see. You write a balance in moles, plug in flow rates that are in kilograms per hour, and wonder why the numbers don't make sense. Convert everything to the same basis before you start writing equations. Assuming constant molar overflow in distillation when the heat of mixing is significant. The constant molar overflow assumption simplifies the McCabe-Thiele method and is fine for ideal or near-ideal systems. For systems with large heat effects like acetone-chloroform or ethanol-water near certain compositions, it introduces real errors. I've seen designs where assuming constant molar overflow led to specifying eight fewer trays than actually needed, which showed up as a purity shortfall during commissioning. Neglecting the mass of catalyst or solid inventory in a reactor balance. Fixed bed reactors hold a significant mass of catalyst. Fluidized beds hold even more. If you're doing a transient balance around a reactor, that inventory matters. Steady-state balances are usually done on the fluid streams only, but if your problem involves startup, shutdown, or catalyst addition, the solid phase is part of your system boundary.
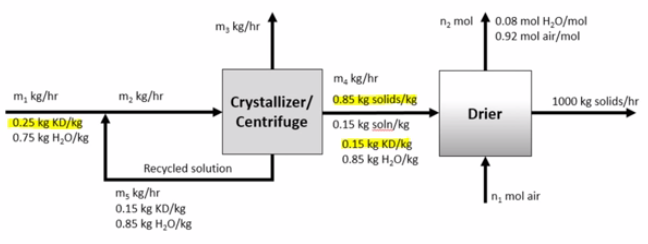
Not checking your balance closure. After you solve, add up all the input streams and all the output streams. If they don't match within your specified tolerance, something is wrong. A ten-to-one percent imbalance is a red flag. A sub-percent imbalance on a well-posed problem is acceptable. The tolerance depends on the application. For process design, one percent is standard. For environmental compliance calculations, you might need much tighter closure because small errors compound across multiple units.
When Material Balances Fail Completely
They fail when the system isn't closed and mass is crossing your boundary in ways you didn't account for. Leaks. Evaporation from open tanks. Absorption into packing material. Sorption onto catalyst surfaces. I worked on a solvent recovery unit where the material balance was consistently off by three percent on the light ends. We checked every stream, recalibrated every analyzer, redid the balance three times. The missing three percent was evaporating from an open flare header that wasn't on the original P&ID. It had been added during a process modification years earlier and nobody had updated the documentation. They also fail when the chemistry is wrong. If your stoichiometry is incorrect or you're missing a side reaction, the balance will close numerically but the results won't match reality. This is harder to catch because the numbers look right. Cross-check with experimental data whenever possible. A single measurement of an outlet composition can tell you whether your reaction model is reasonable or completely off.
Practical Workflow for a Typical Problem
Draw the process diagram with all known flow rates, compositions, temperatures, and pressures. Mark the system boundaries. Label every unknown. Do the degrees of freedom analysis. If the problem isn't solvable, identify what additional information you need. Solve the non-reactive parts first, then the reactive sections. Handle recycle loops with iteration. Check your closure. If it doesn't close, trace back through your equations to find where the error entered. This workflow usually takes twenty to forty minutes for a single-unit steady-state problem with five to ten streams. A multi-unit system with recycle and reactions might take two to four hours depending on complexity. Using a process simulator cuts that down significantly, but only if you set it up correctly, which brings us back to understanding the fundamentals rather than treating the software as a black box.