Understanding how electrons actually sit in molecules
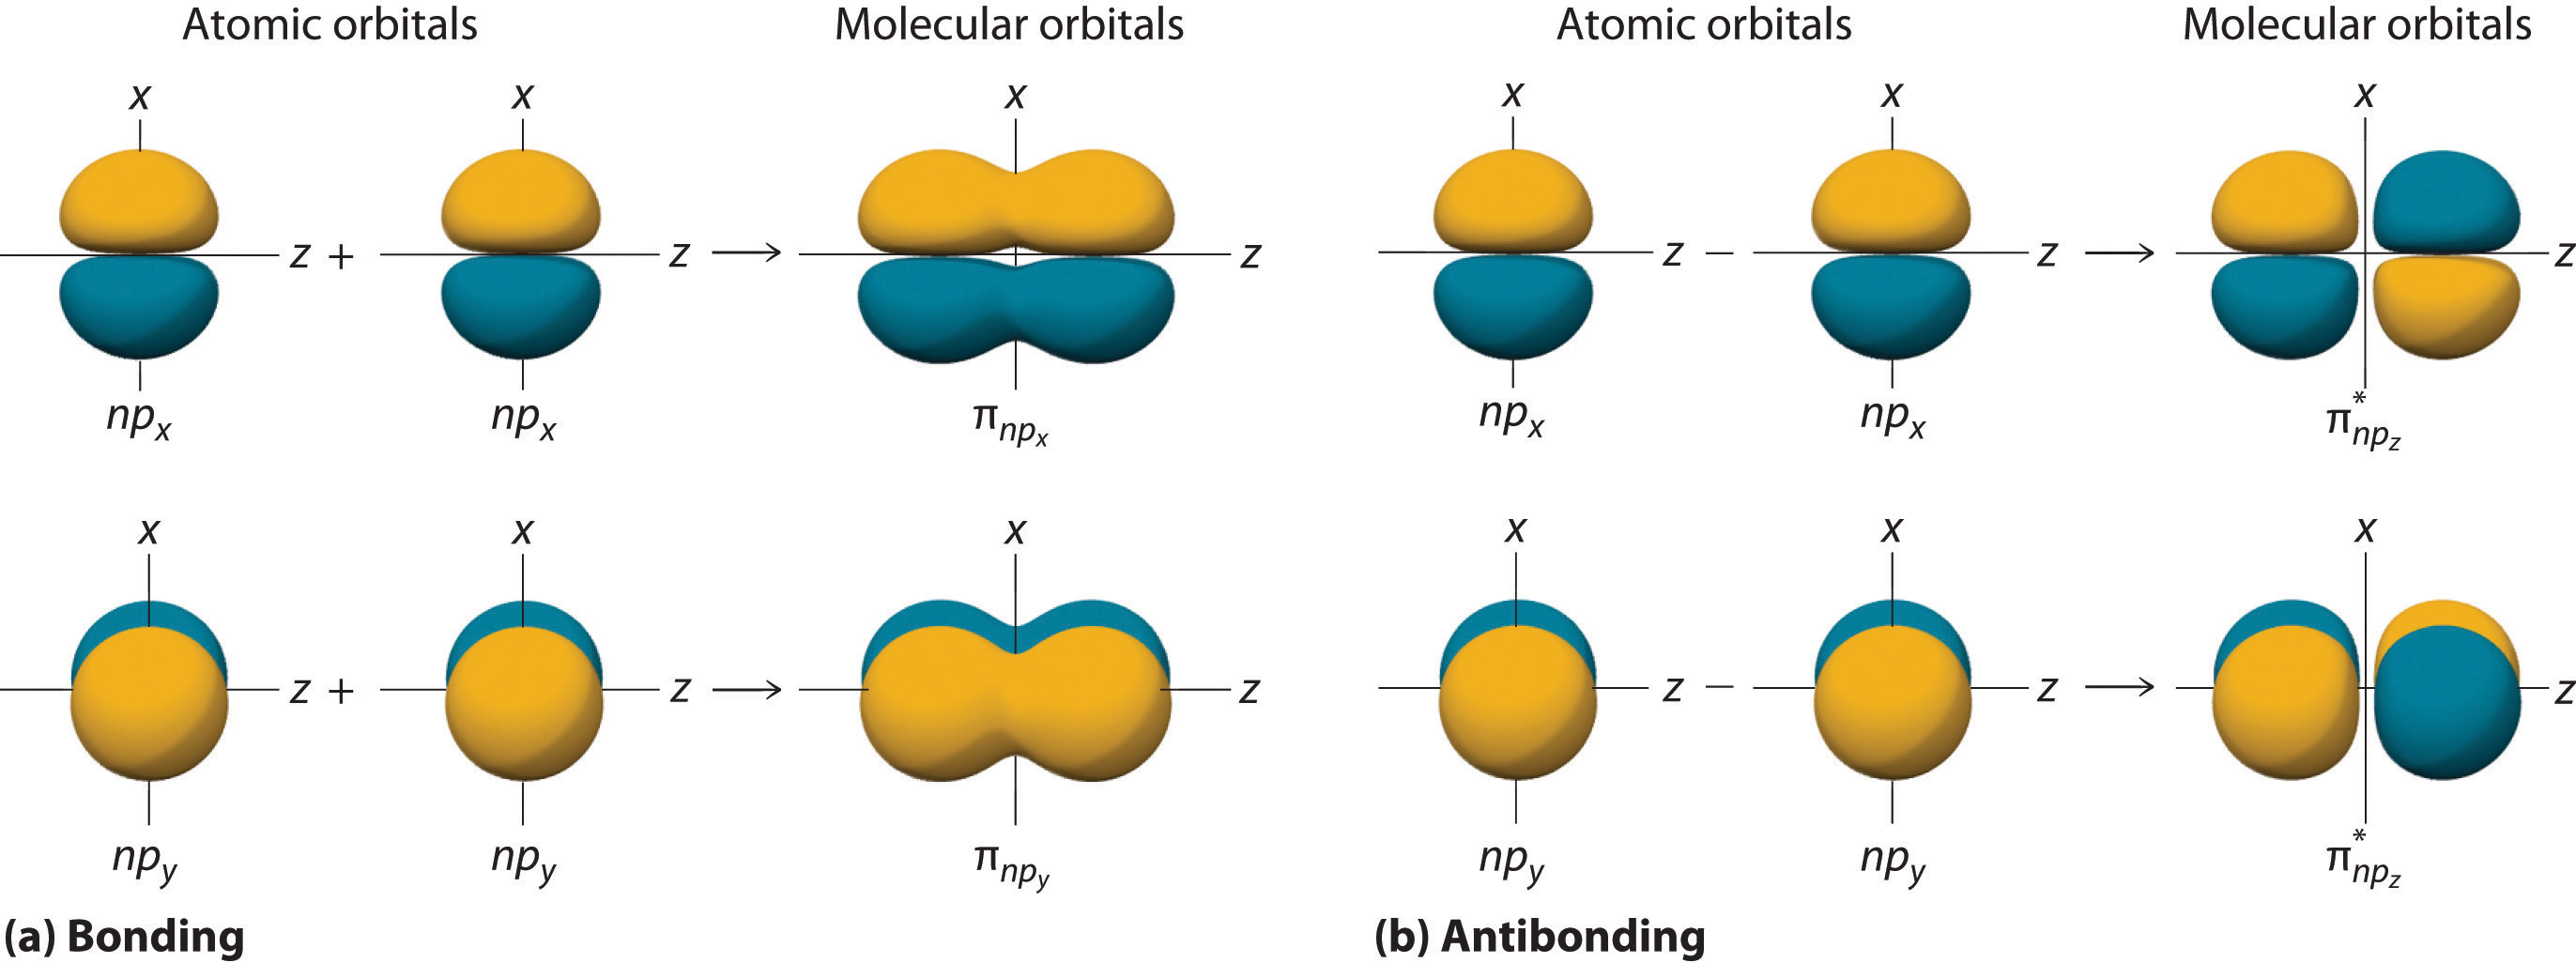
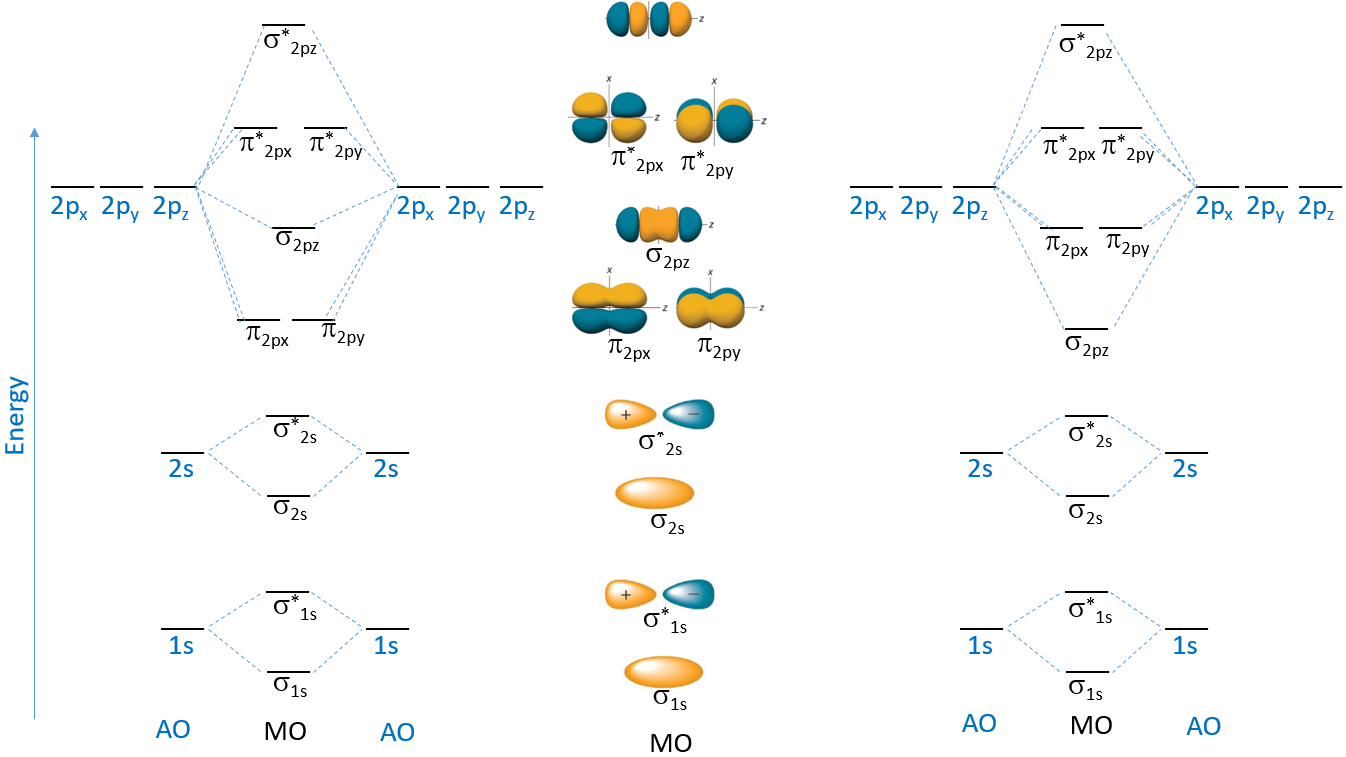
Molecular orbital theory is the standard way chemists describe where electrons live when atoms bond together. Forget the old picture of electrons dancing between two nuclei like a spring. In reality, each electron occupies a molecular orbital that belongs to the whole molecule, and those orbitals stretch across all the atoms involved. First you draw the atomic orbitals of each atom on the sides of a diagram. Then you draw the molecular orbitals in the middle. The number of molecular orbitals always equals the number of atomic orbitals you started with. That is a strict rule, not a suggestion. Electrons fill from the lowest energy upward. You follow the Aufbau principle, the Pauli exclusion principle, and Hund's rule, exactly like you do for atoms. But here is the practical part that most textbooks skim over: the ordering of the molecular orbitals changes depending on which atoms you are dealing with. For B2, C2, and N2, the 2p orbital sits higher in energy than the 2p orbitals. For O2, F2, and Ne2, the order flips and 2p drops below 2p. If you get that wrong, your bond order calculation is wrong, and your prediction about magnetism is wrong too.
I learned this the hard way when I was grading first-year lab reports. Every single student drew the O2 diagram using the N2 ordering. They put six electrons in 2p before the 2p, which made O2 diamagnetic. Oxygen is not diamagnetic. It is paramagnetic, and anyone who has run a simple magnet experiment knows that. The correct configuration places the last two electrons singly in the two degenerate *2p orbitals. That is the classic proof that MO theory actually works.
Reading the diagram and extracting useful numbers
Once the diagram is drawn correctly, you count bonding electrons and antibonding electrons. The bond order formula is straightforward: bond order equals the number of bonding electrons minus the number of antibonding electrons, divided by two. A bond order of two means a double bond. A bond order of one point five means something in between a single and a double bond, like the superoxide ion O2-. You can also predict stability. If the bond order drops to zero or below, the molecule does not exist under normal conditions. That is why He2 is not a stable species, even though it occasionally shows up in mass spectrometry at very low pressures. For heteronuclear diatomics like CO or NO, the orbitals are not symmetric around the center. The more electronegative atom pulls the bonding orbitals closer to its own atomic energy levels, and the antibonding orbitals sit closer to the less electronegative atom. This matters when you are trying to understand where the highest occupied molecular orbital is located, because that determines which atom tends to donate electrons in a reaction.
Get the Full Details

Common pitfalls and the edge cases that trip people up
The biggest mistake is assuming every diatomic molecule follows the same orbital diagram. The s-p mixing effect is real and significant for B2 through N2, but it becomes negligible once you reach O2 and beyond. I once spent an afternoon debugging a computational chemistry script because I had hardcoded the N2 orbital ordering for every molecule it processed. The output was garbage for O2, F2, and everything else. The fix was simple: add a conditional that switches the 2p and 2p ordering based on the total number of valence electrons. Anything above fourteen valence electrons gets the flipped ordering. Another subtle issue is treating molecular orbitals as strictly localized. In polyatomic molecules, the concept of bonding versus antibonding becomes fuzzier. Delocalized orbitals span three or more atoms, and drawing them by hand is nearly impossible for anything beyond simple cases like benzene or ozone. In practice, you switch to computational software after about four atoms. Gaussian, ORCA, or even free tools like Avogadro will generate the orbitals for you, but you still need to understand the theory to interpret the output correctly.
When MO theory falls short
MO theory is powerful, but it is not a silver bullet. For large organic molecules, the number of molecular orbitals grows rapidly, and the diagram becomes a wall of lines that tells you very little without computation. Transition metal complexes introduce d-orbital splitting that requires crystal field theory or ligand field theory to make sense of, because the simple diatomic MO diagram does not scale cleanly to octahedral or tetrahedral geometries. And for metallic bonding, you end up with bands rather than discrete orbitals, which is a different framework altogether. If you are only interested in bond lengths and basic reactivity, valence bond theory with resonance structures might be faster. MO theory shines when you need to explain magnetism, electronic spectra, or why certain molecules conduct electricity while others do not. Those are the scenarios where the Molecular Orbital Electron Configuration actually earns its keep.