CO Bonding and Why It Doesn't Look Like What You Expect
The molecular orbital diagram for carbon monoxide is one of those topics where every textbook gets slightly different from the next, and most of them miss the part that actually matters for anyone doing real computational work or interpreting spectroscopic data. The basics are straightforward enough. You have ten valence electrons total—four from carbon, six from oxygen—and you're filling bonding and nonbonding orbitals in a way that produces a bond order of three. That's the summary version. The actual shape of the diagram depends heavily on how you handle the energy ordering of the sigma and pi systems, and that's where people start arguing in comments sections.
Molecular Orbital Structure Of Co: The Actual Diagram
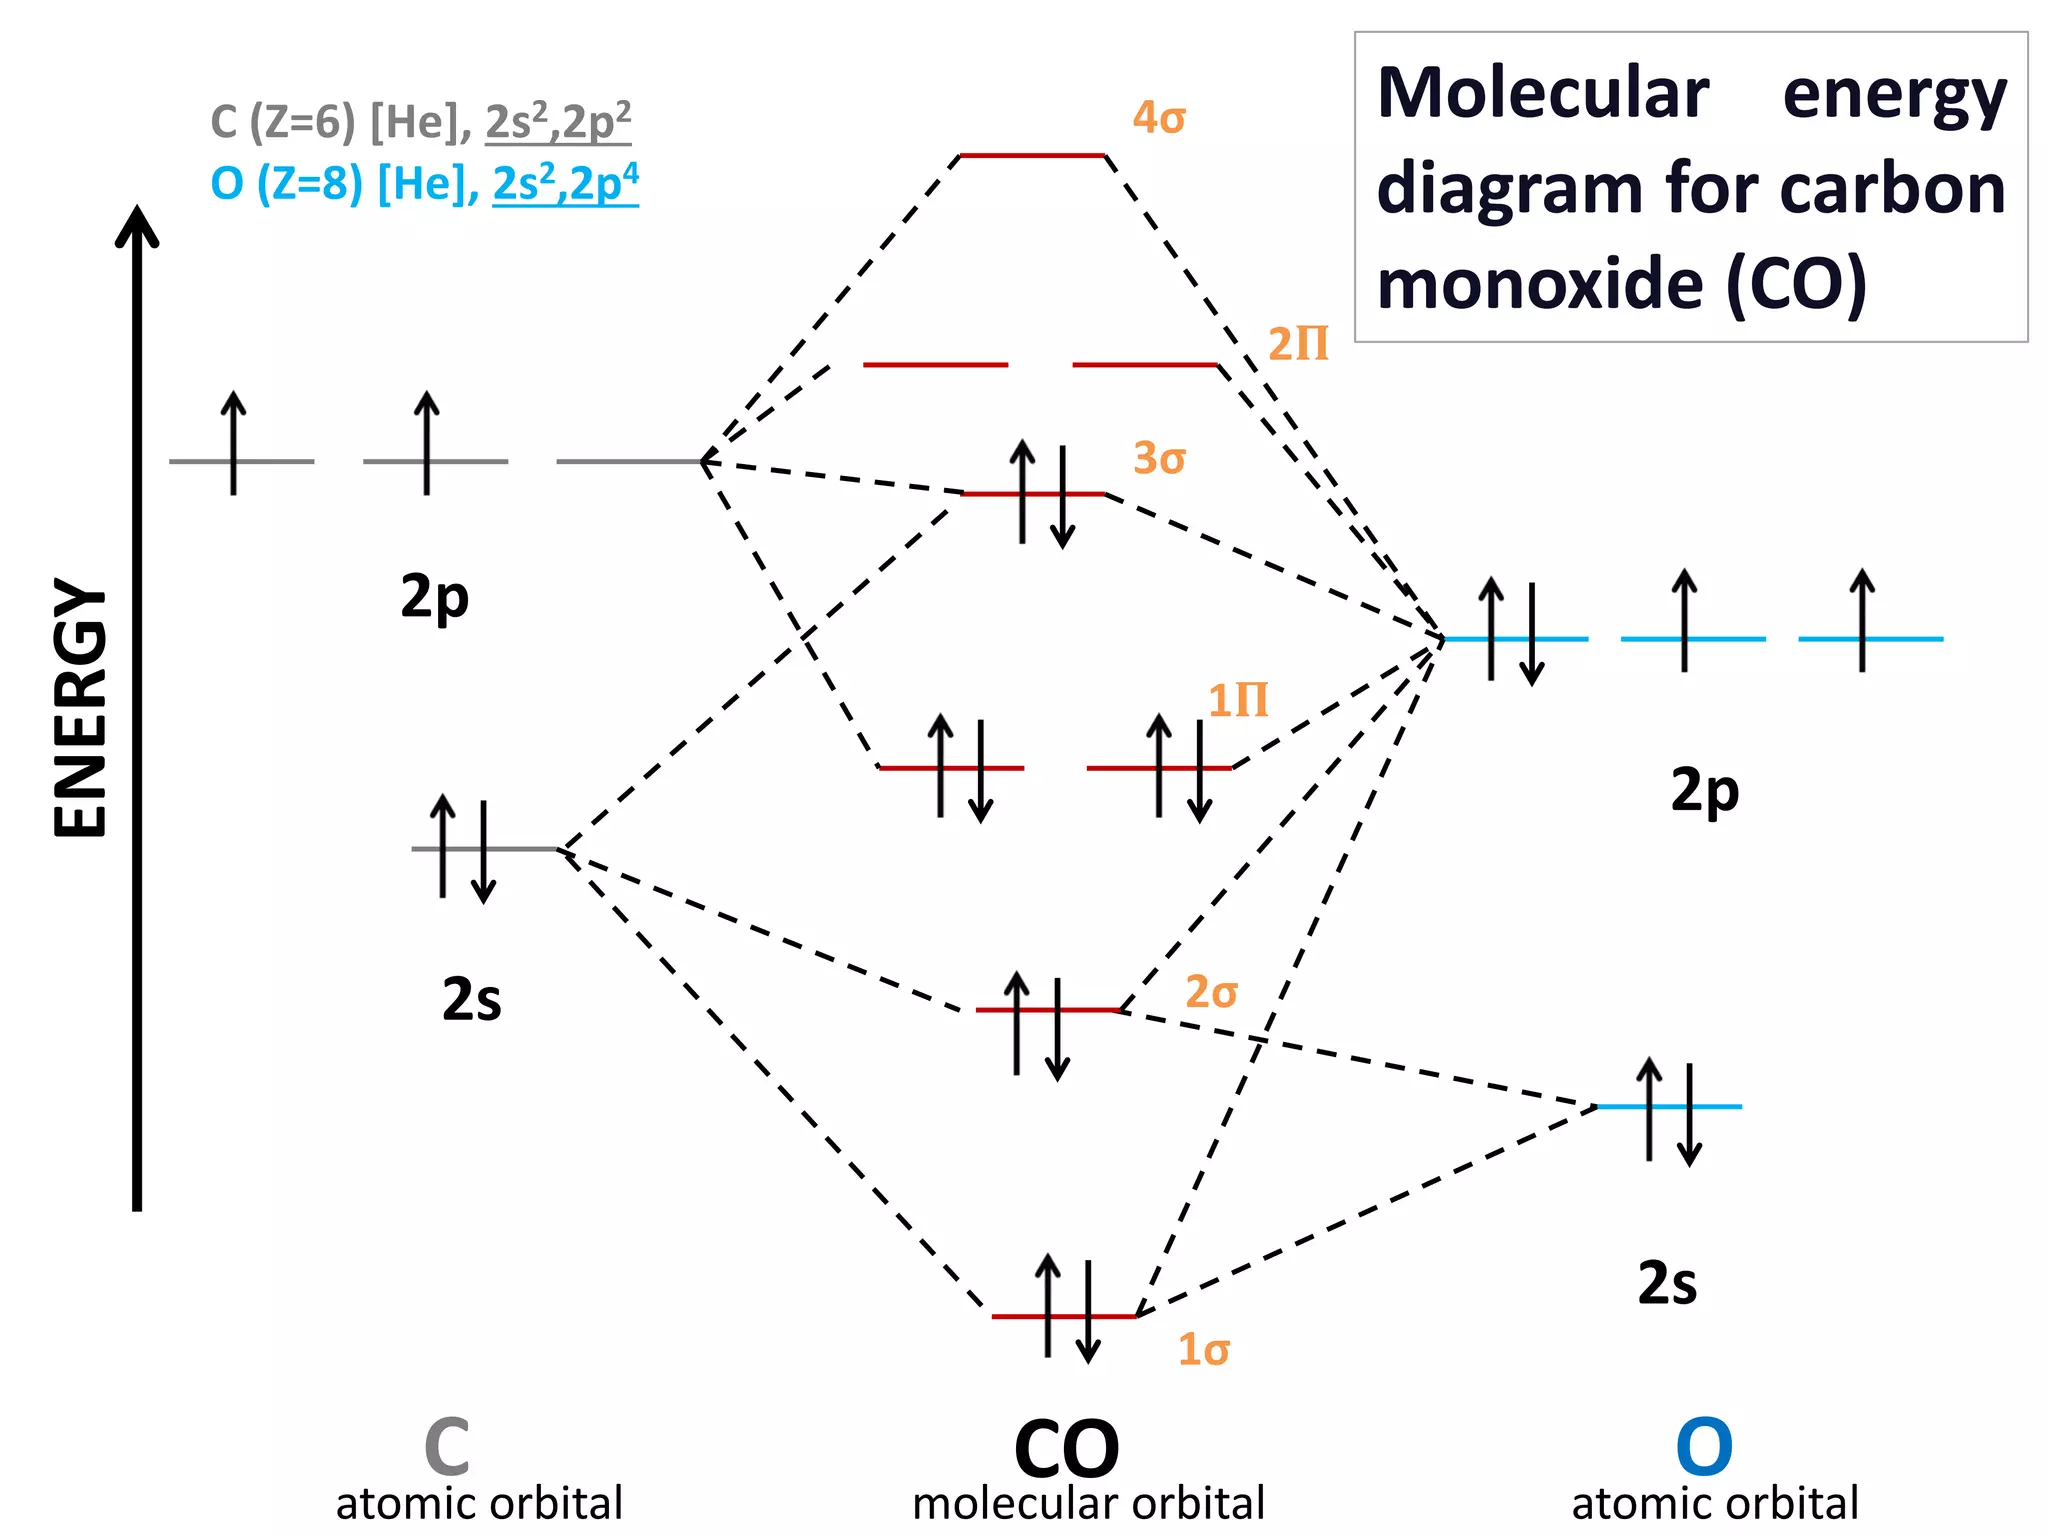
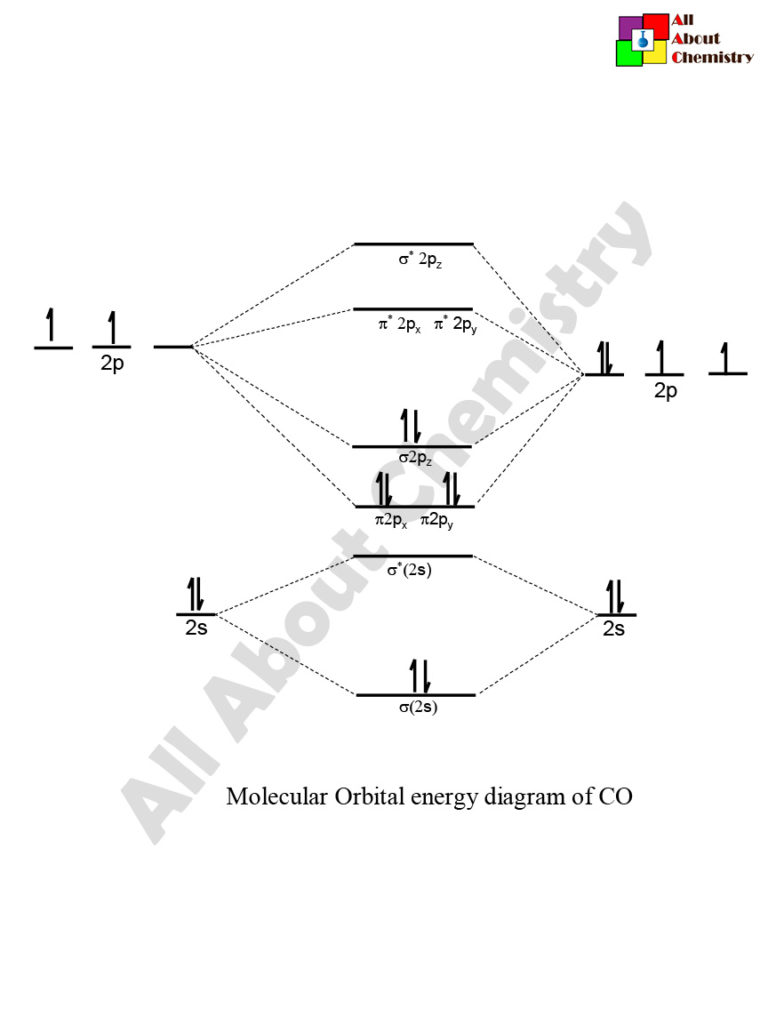
Start with the 2s and 2p atomic orbitals from both atoms. Oxygen's are lower in energy than carbon's because oxygen is more electronegative, and that asymmetry does most of the heavy lifting. The resulting MOs, going from lowest to highest energy, run like this: sigma 2s bonding, sigma 2s antibonding, sigma 2p bonding, two degenerate pi 2p bonding orbitals, two degenerate pi 2p antibonding orbitals, and then the sigma 2p antibonding orbital at the top. You fill ten electrons into that framework, giving you a configuration of (2s)²(*2s)²(2p)²(2p), which leaves the antibonding and orbitals empty. The bond order calculation is (8 bonding minus 2 antibonding) divided by 2, which gives you exactly 3. That's why CO is so stable and why it binds to transition metals through the carbon end rather than the oxygen end—the highest occupied molecular orbital, the HOMO, is the 2p bonding orbital with its electron density concentrated more on the carbon atom. Here's the thing most undergraduate textbooks gloss over. The 2s and *2s orbitals aren't purely bonding and antibonding in any clean sense. There's significant mixing between the 2s and 2p_z orbitals on both atoms, and depending on your computational method, that mixing can shift the energy ordering enough that the 2p drops below the 2p or vice versa. The experimental photoelectron spectrum settles this for CO, and it shows the orbitals coming from lone pairs at around 14 eV and 17 eV ionization energy, while the bonding orbitals sit deeper at about 16 eV. The exact ordering of those three filled and levels is sensitive to the method you use, and if you're running DFT calculations on CO complexes, you will see different orderings depending on whether you're using B3LYP, PBE, or a hybrid functional with different exact exchange percentages.
I spent a couple of weeks last year trying to reconcile computed CO stretching frequencies with infrared spectra for a nickel carbonyl complex, and the problem kept coming back to how the functional handled the CO backbonding. Standard B3LYP was overestimating the backdonation from nickel into the CO * orbital, which weakened the C-O bond too much in the calculation. The frequency dropped to around 1950 cm¹ instead of the experimental 2056 cm¹. The fix wasn't elegant. I switched to B97X-D with a triple-zeta basis set, which gave better treatment of the long-range exchange and cut the error down to about 30 cm¹. If you're working with metal carbonyls, don't trust a single functional to get this right without benchmarking against a known reference first.
Get the Full Details

What the LUMO Actually Means
The lowest unoccupied molecular orbital in CO is the * antibonding orbital, and it's doubly degenerate. That's the orbital that accepts electron density during backbonding from a transition metal, and it's also the orbital involved in most of the photochemistry around CO. When CO gets excited by UV light, an electron jumps from the bonding orbital into the * antibonding orbital, which weakens the bond and makes the molecule far more reactive. This is relevant if you're doing anything with photodissociation studies or mass spectrometry of metal carbonyl clusters. The dipole moment of CO is another point where the simple Lewis structure lies to you. Carbon monoxide has a very small dipole moment of about 0.12 D, and it points in the wrong direction compared to what you'd predict from electronegativity alone. Oxygen should be the negative end, but the dipole actually points toward the oxygen being slightly positive relative to carbon. The molecular orbital picture explains this through the lone pair on carbon and the asymmetry in orbital mixing, but honestly the simplest explanation is that the HOMO is carbon-centered and the electron density distribution doesn't match what a naive formal charge model would suggest. If you're building force fields or doing molecular dynamics with CO, that tiny dipole moment is a practical problem. Most general force fields parameterize CO with a point charge on each atom, and getting the electrostatic potential right requires putting a partial positive charge on oxygen and a partial negative on carbon, which feels counterintuitive but matches the actual electron density. I've seen people use charges fromRESP fitting or Mulliken population analysis and get completely wrong solvation behavior because the raw charges don't capture the anisotropy of the electron distribution.
Common Mistakes and Where the Model Breaks
The biggest mistake people make with the MO diagram of CO is treating it as static. The diagram you draw on paper assumes the molecule is isolated and at equilibrium geometry. The moment you coordinate CO to a metal, the orbital energies shift substantially. The donation from carbon to the metal raises the energy of the orbital, and the backdonation lowers the energy of the * orbital. Both effects strengthen the metal-carbon bond and weaken the C-O bond, which is why IR stretching frequencies drop from 2143 cm¹ in free CO to somewhere between 1850 and 2100 cm¹ in metal carbonyls depending on the oxidation state and coordination environment. Another thing to watch out for is the assumption that the bond order of 3 means CO can't do much else. The molecule participates in a surprising number of reactions precisely because the * LUMO is low-lying enough to accept electrons, and the HOMO is high enough to donate them. In organometallic chemistry, CO acts as both a sigma donor and a pi acceptor, and the balance between those two roles determines the reactivity of the complex. For example, in Wilkinson's catalyst or in cobalt-catalyzed hydroformylation, the CO ligand can be displaced or inserted depending on the electronic properties of the other ligands on the metal. The MO picture also breaks down when you get to larger clusters or surface-bound CO. On a platinum surface, for instance, the CO 5 and 1 orbitals hybridize with the metal d-band, and you're no longer dealing with discrete molecular orbitals. You get resonance widths and adsorption geometries that depend on whether the CO is sitting on top, in a bridge position, or in an hollow site. The stretching frequency shifts by hundreds of wavenumbers depending on the adsorption site, and a simple two-atom MO diagram won't predict any of that.
If you need accurate orbital energies for CO, DFT is convenient but not particularly reliable without careful validation. For single-reference cases, coupled-cluster methods like CCSD(T) with a large basis set will give you something close to the experimental photoelectron spectrum, but they're expensive and scale poorly if you're doing anything beyond small systems. The standard approach in the literature is to use DFT for geometry and trends, then validate key orbital energies against higher-level calculations or experimental data before drawing conclusions about reactivity or spectroscopy.