Ammonia Geometry, Actually
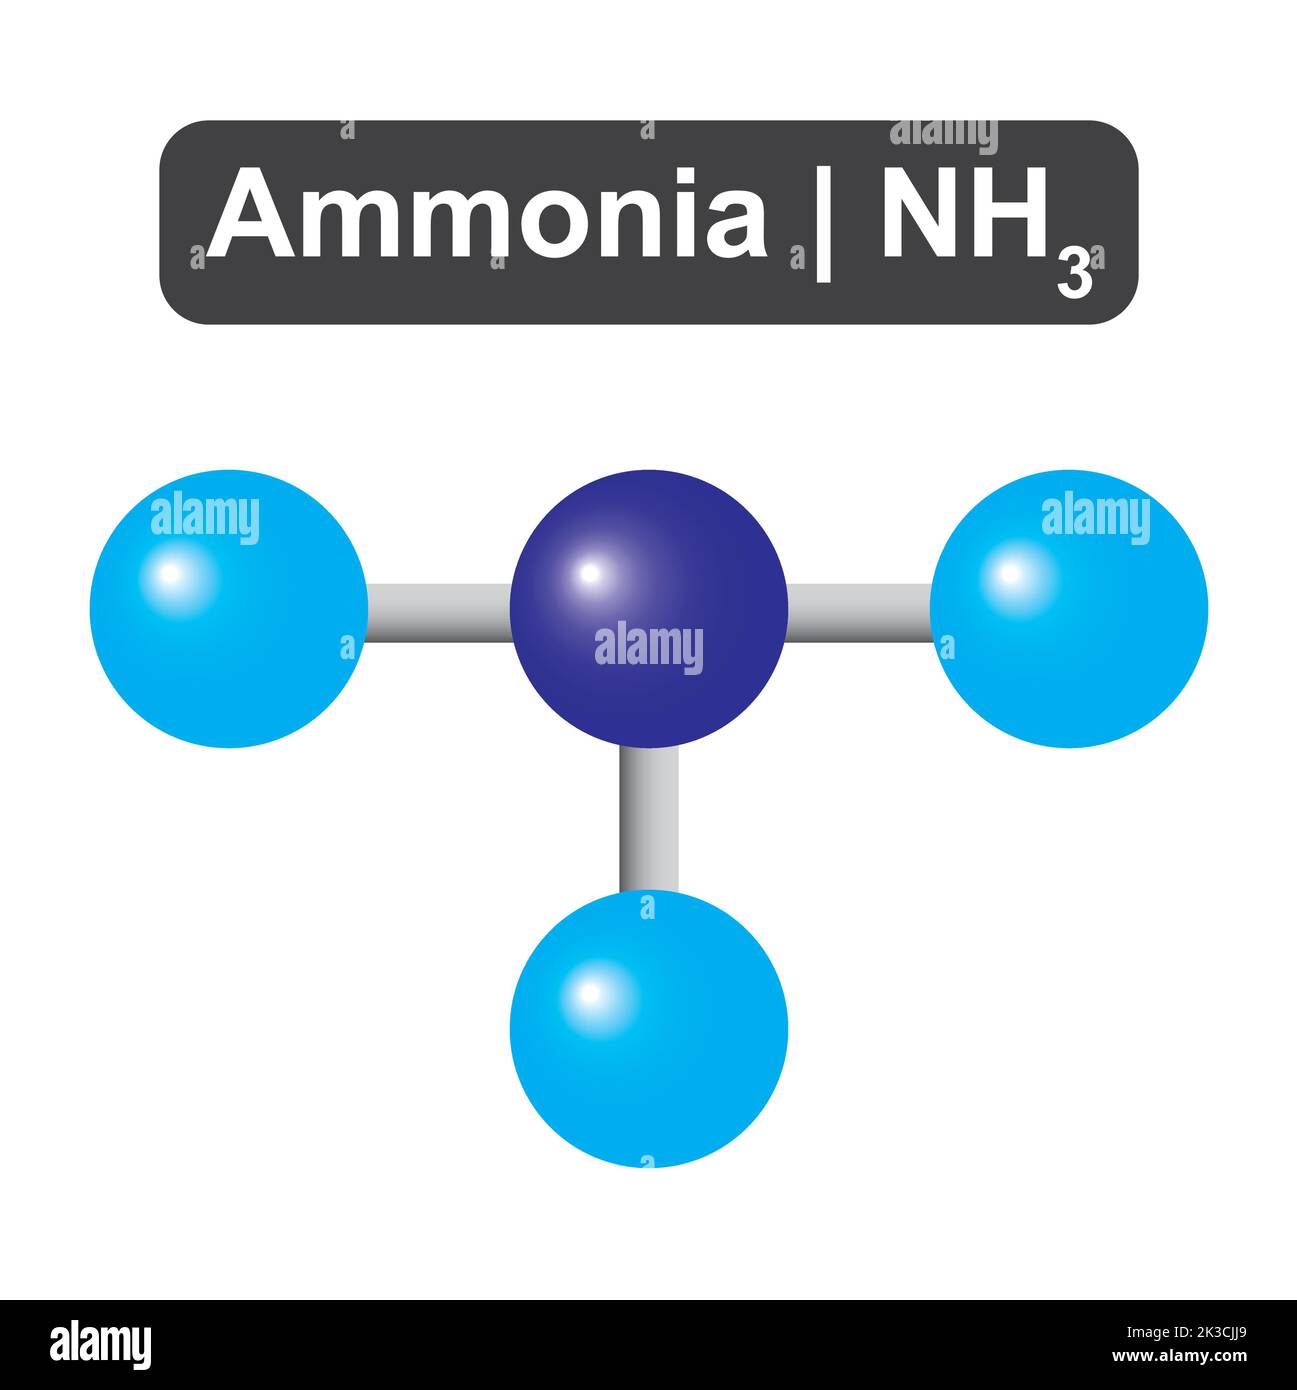
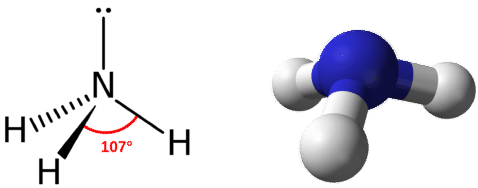
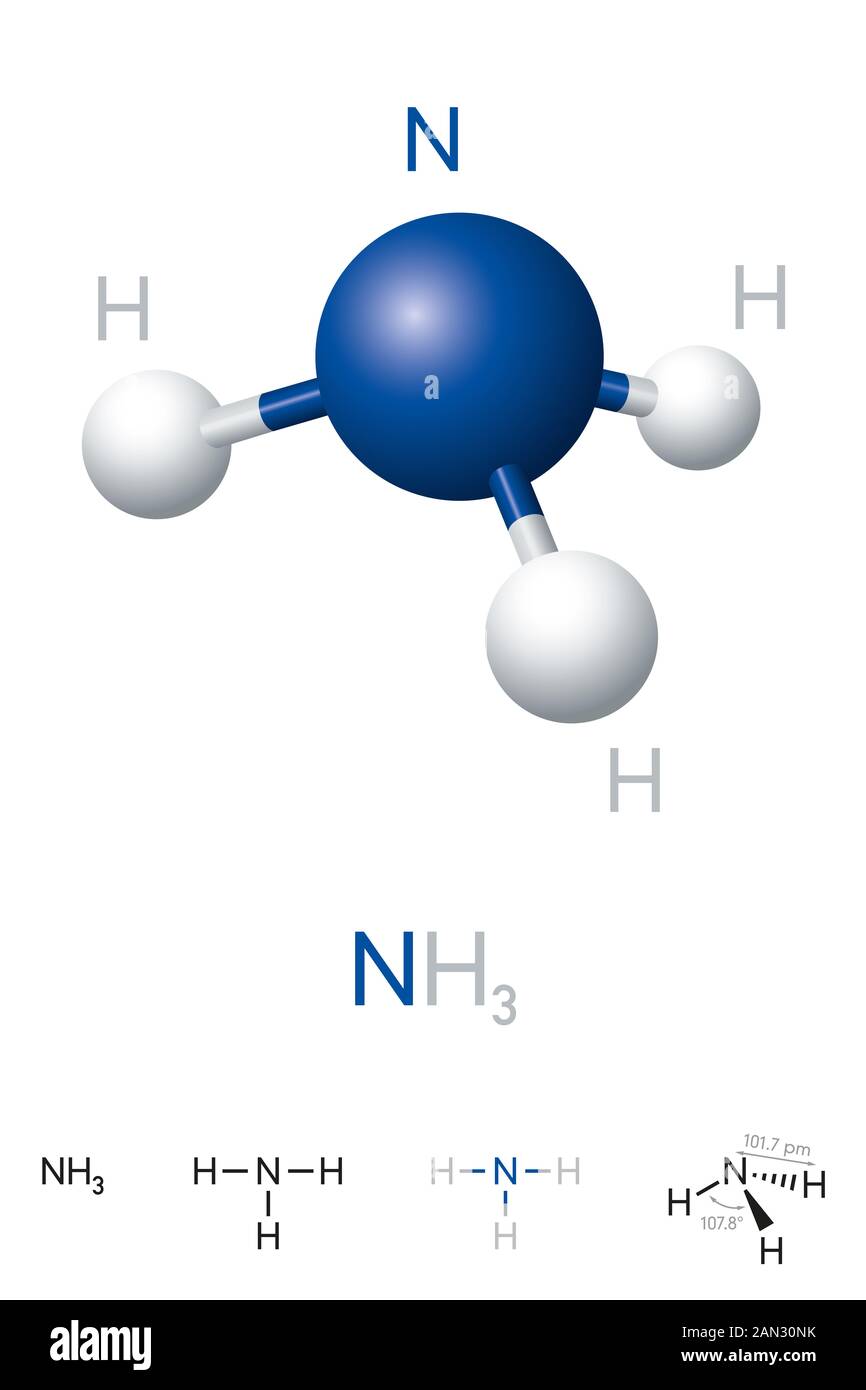
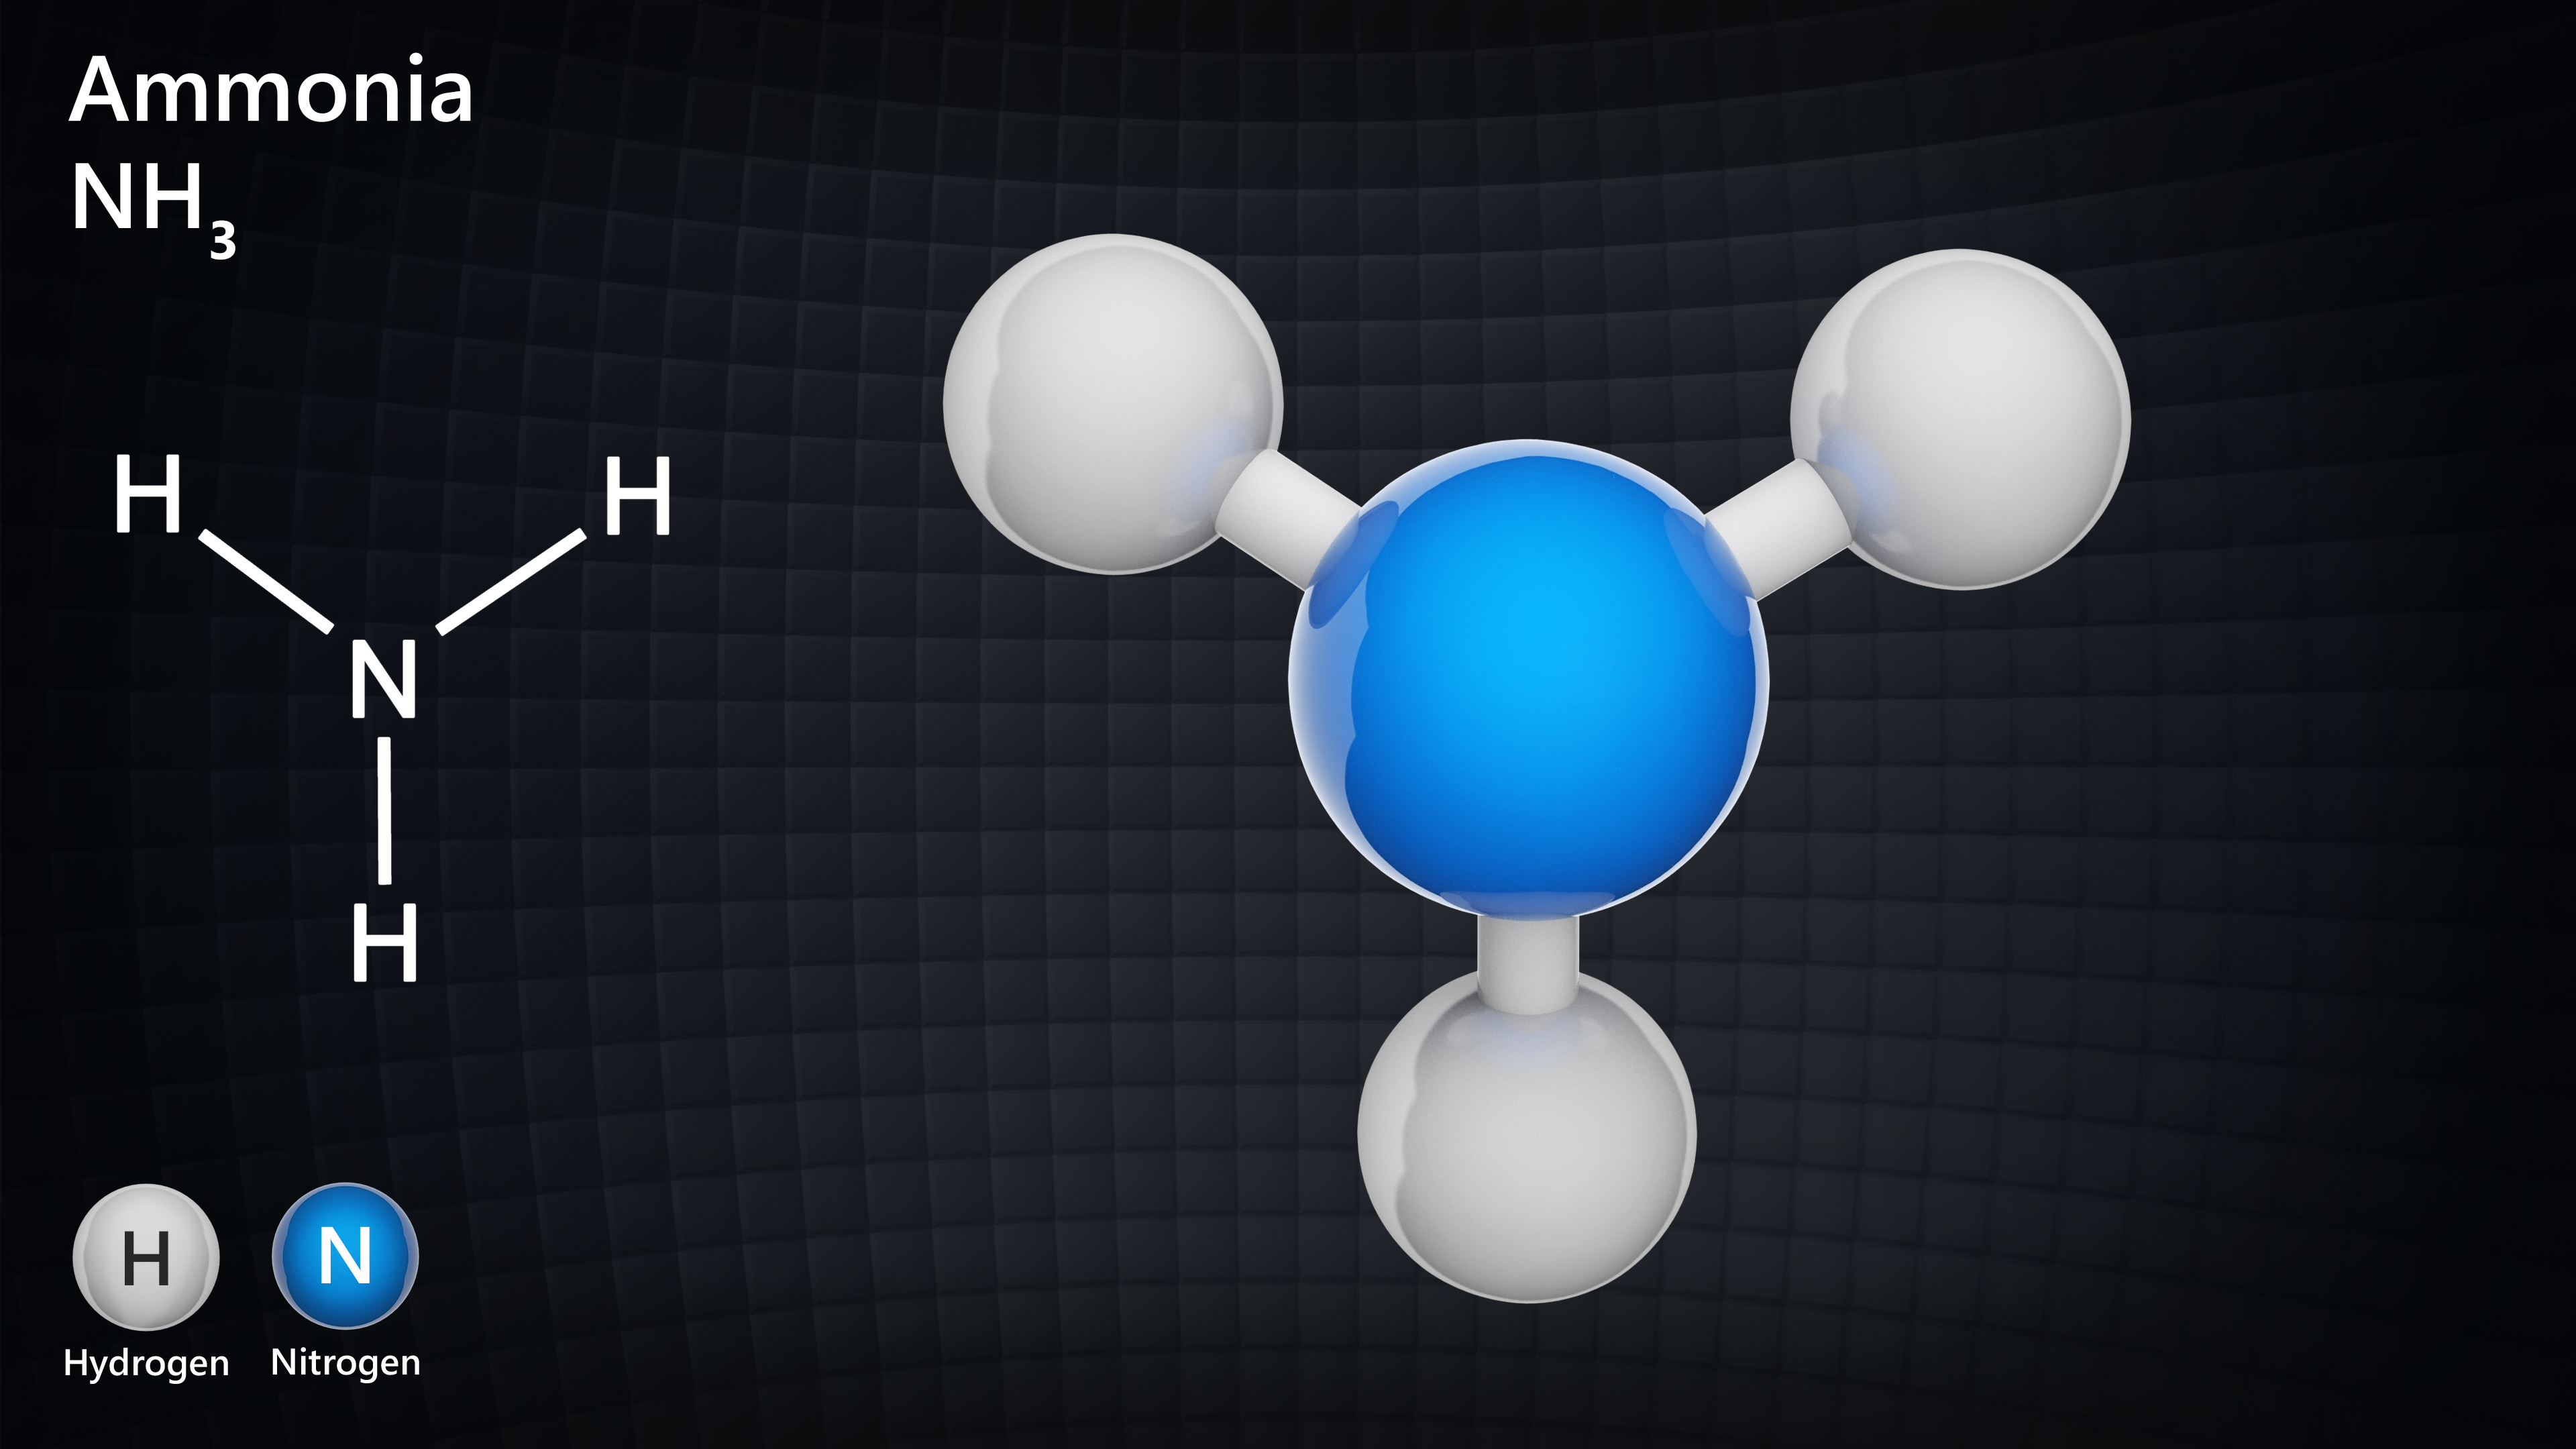
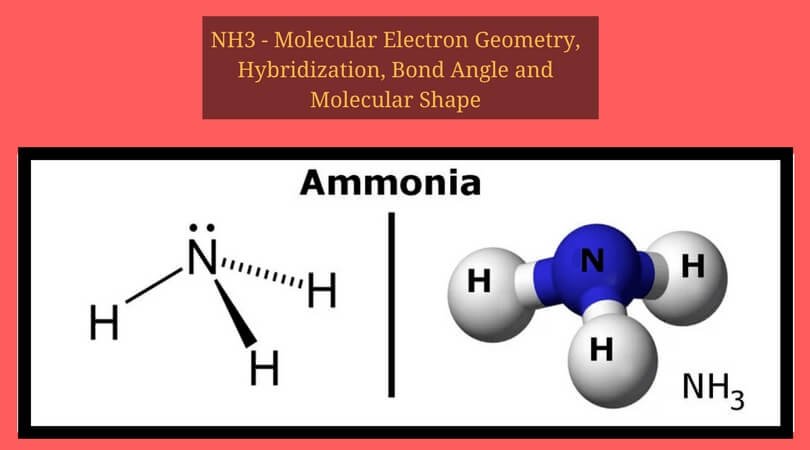
The central nitrogen in NH3 has three bonding pairs and one lone pair, which means the atom itself adopts a trigonal pyramidal arrangement rather than anything flat or symmetrical. The three hydrogens form the base of a pyramid with nitrogen sitting above the plane they define. This comes from the underlying tetrahedral electron geometry, where the four electron domains around nitrogen spread out as far as possible. The molecular shape only considers atom positions though, so that lone pair disappears from the final description even though it is actively pushing the N-H bonds downward. I keep running into grad students who conflate electron geometry with molecular geometry, and it costs them points on exams or makes them misread spectroscopic data later. The distinction matters because X-ray diffraction and electron diffraction experiments detect nuclei, not lone pairs. When someone reports the shape of ammonia from an experiment, they are describing the three hydrogen positions relative to nitrogen, not the full tetrahedral electron distribution. That is why the accepted value sits at approximately 107 degrees for the H-N-H bond angle instead of the ideal 109.5 degree tetrahedral angle. The lone pair occupies more spatial volume than a bonding pair and compresses the adjacent bonds slightly. The VSEPR framework gets you most of the way there without running any computations. Count valence electrons: nitrogen contributes five, each hydrogen contributes one, giving eight total. Subtract six electrons used in the three N-H bonds and you are left with a lone pair on nitrogen. Four electron domains means sp3 hybridization on paper, tetrahedral electron geometry, and trigonal pyramidal molecular geometry as the observable shape. This works reliably for ammonia and similar molecules, but it breaks down quickly when you get into transition metal complexes or molecules with significant d-orbital contribution, which is not rare in coordination chemistry.
I had a specific issue last year where a student was trying to reproduce ammonia geometry in a Gaussian calculation and kept getting distorted results. The problem was not the theory, it was the initial guess geometry. I had placed the hydrogens in a planar arrangement before running the optimization, and the single-point energy calculation never converged to the correct pyramidal minimum. The workaround was straightforward: manually position nitrogen slightly above the hydrogen plane by about 0.4 angstroms before running the geometry optimization, which gives the optimizer a clear pathway to the correct local minimum. This kind of bad initial guess is something standard quantum chemistry packages do not always handle gracefully, especially with tighter convergence criteria. Bond lengths in NH3 are experimentally around 1.012 angstroms for each N-H bond, and the bond angle is 107.8 degrees from microwave spectroscopy data. If you need a quick reference structure, the CCCBDB database at NIST lists these values along with uncertainty estimates, which is more reliable than most textbooks that round aggressively. Computational chemists often cite the same numbers when validating new functionals against benchmark data.
Practical Modeling Approaches
When I need to visualize or manipulate the ammonia geometry quickly, I reach for a molecular modeling kit first because there is something about physically holding the atoms that resolves confusion faster than any diagram. A standard kit with the correct connectors shows immediately why ammonia cannot be planar, and why the lone pair region looks larger than the bonding regions. After that, I sometimes load the coordinates into Avogadro or Mercury to rotate the structure and check dihedral angles, though for a single small molecule this is overkill. The structure is rigid enough that rotation alone tells you everything you need to know. For computational work, a basis set like 6-31G* is adequate for routine geometry optimization of ammonia. If you need high accuracy, particularly for vibration-rotation interactions or precision benchmarks, you should use a larger triple-zeta basis set with polarization functions on hydrogen, such as aug-cc-pVTZ. The difference in computed bond angle between these levels is small, roughly 0.3 to 0.5 degrees, but it matters when you are publishing or comparing against experimental spectra.
Get the Full Details
Where The Simple Model Fails
VSEPR treats ammonia as a simple isolated molecule with localized electron pairs, which works fine for predicting shape but does not capture everything about its behavior. The lone pair on nitrogen is responsible for ammonia acting as a Lewis base, coordinating to metal centers, and participating in hydrogen bonding networks. None of that chemistry is explained by the geometry alone. When ammonia forms hydrogen bonds with water or binds to a metal ion, the local geometry around nitrogen distorts, and the simple VSEPR picture no longer predicts the new arrangement accurately. You need to consider steric effects from ligands, electronic effects from the metal center, and sometimes relativistic effects in heavier analogs like phosphine. Another limitation is that VSEPR does not give you quantitative predictions. It tells you the shape is trigonal pyramidal but not exactly what angle to expect without experimental data or computation. This is fine for most undergraduate courses but insufficient for research where you need precise structural parameters. DFT calculations fill that gap, though they require computational resources and proper validation against known benchmarks. For routine lab work, I usually rely on published crystallographic data from the Cambridge Structural Database when I need exact geometries rather than running my own calculations. Hydrogen bonding complicates things further. In liquid ammonia or aqueous solution, each molecule participates in a dynamic network that transiently distorts the ideal geometry. Neutron diffraction studies show that the H-N-H angle can vary by a degree or two depending on the local hydrogen bonding environment. If you are studying ammonia in condensed phases, the gas-phase geometry is only an approximation, and you should account for environmental effects in your analysis.