Understanding how water molecules actually connect
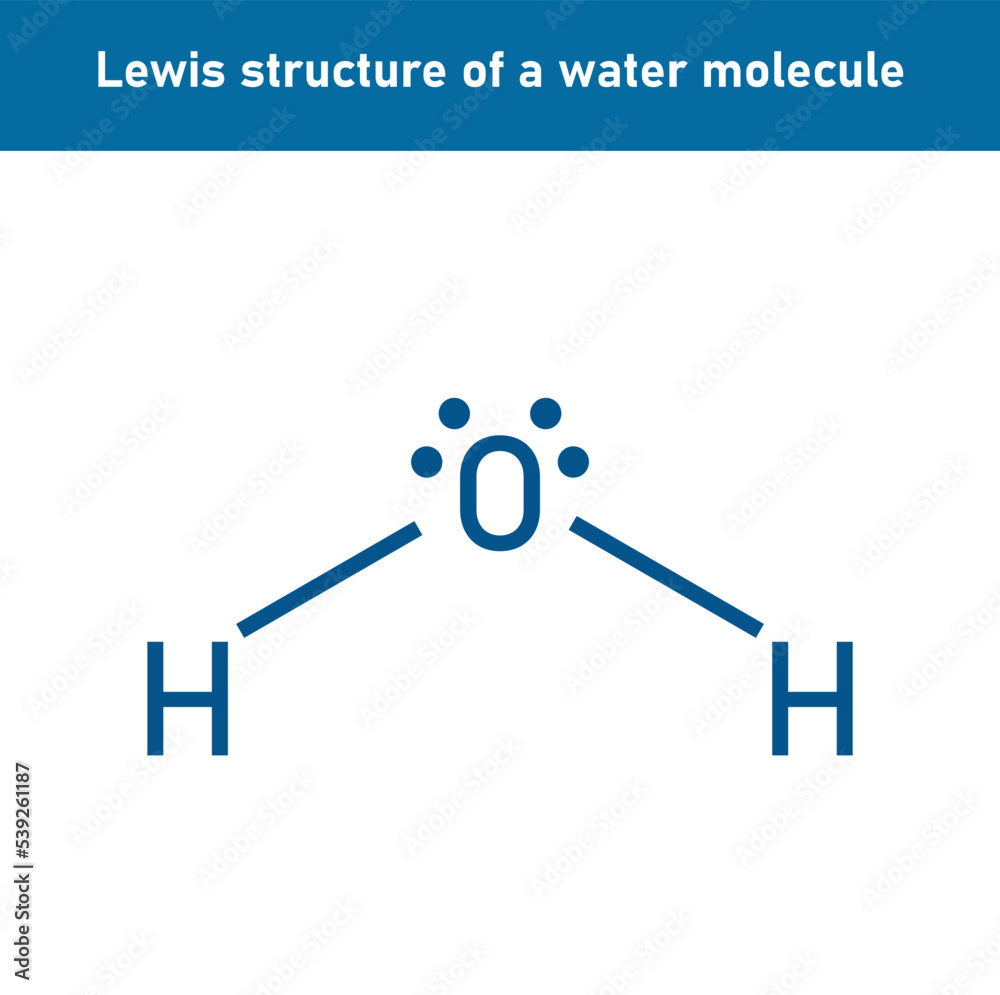
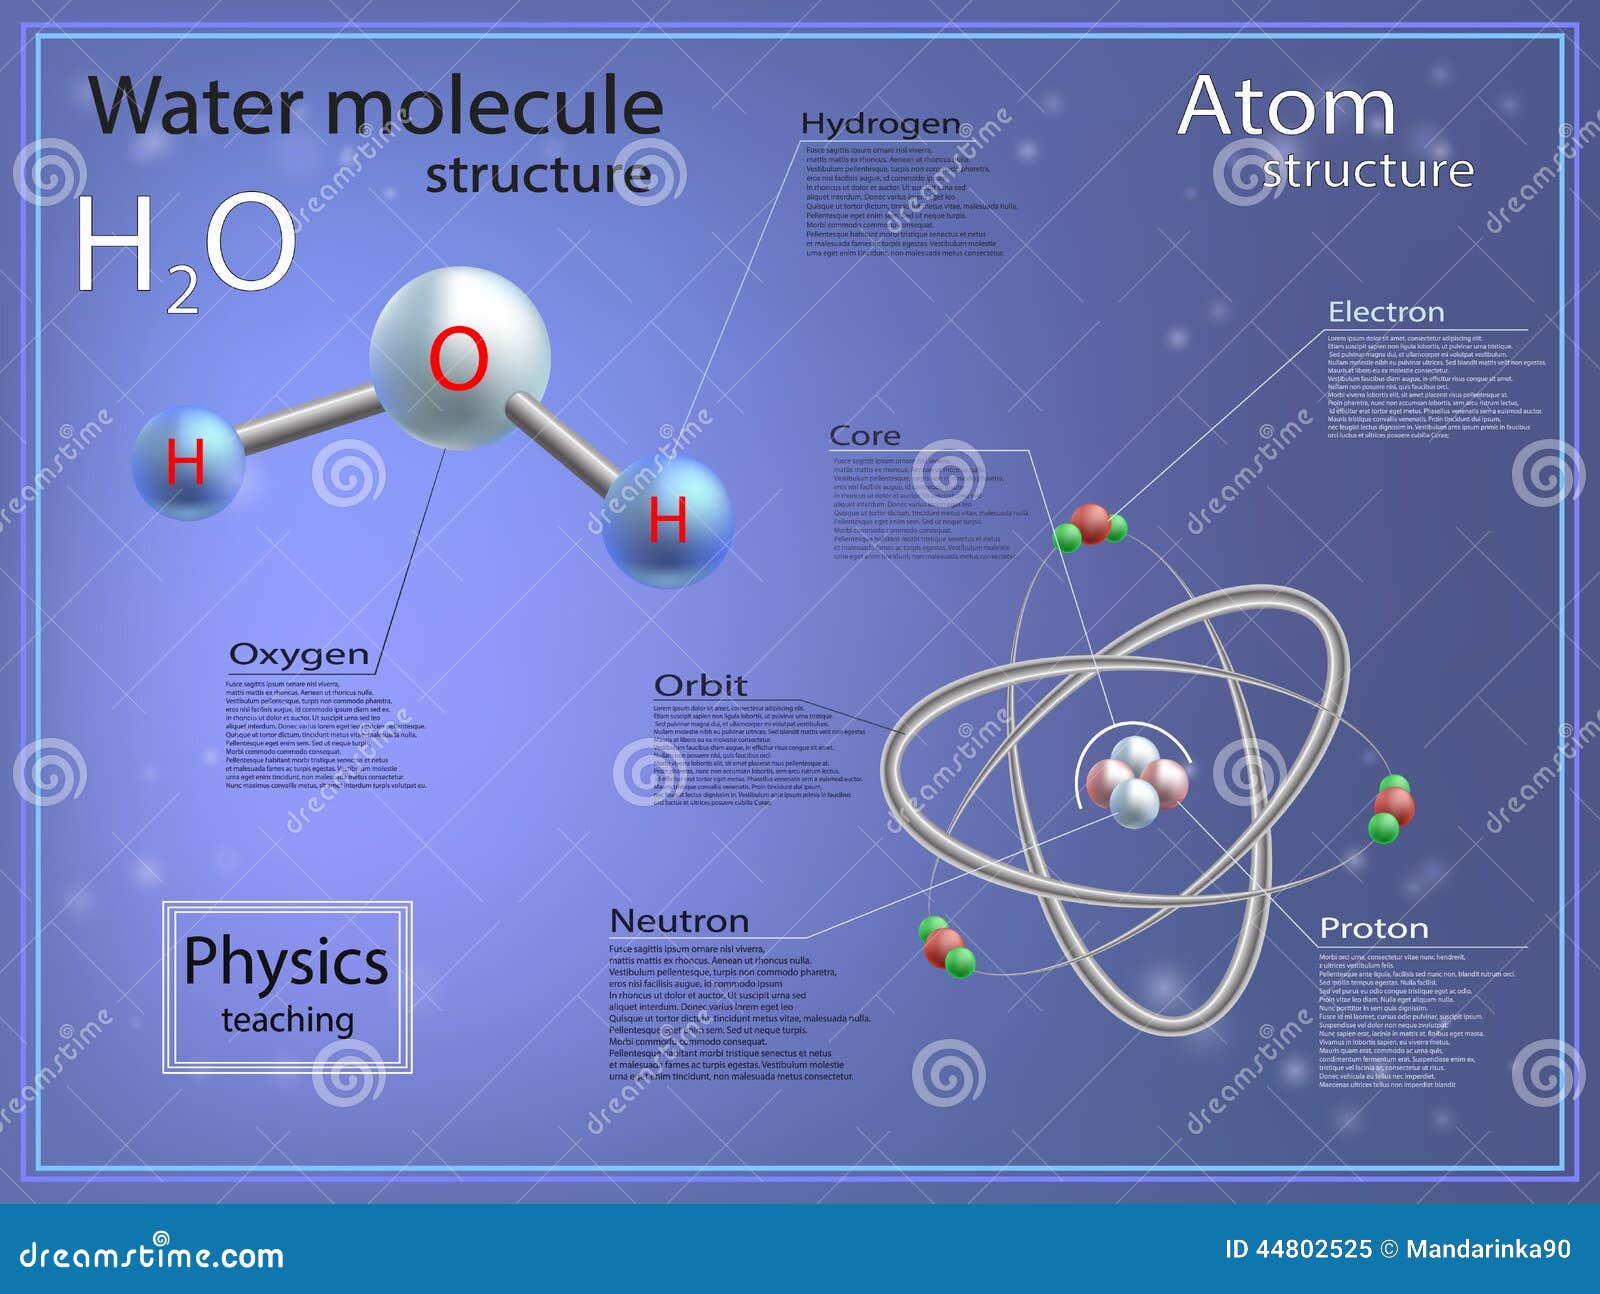
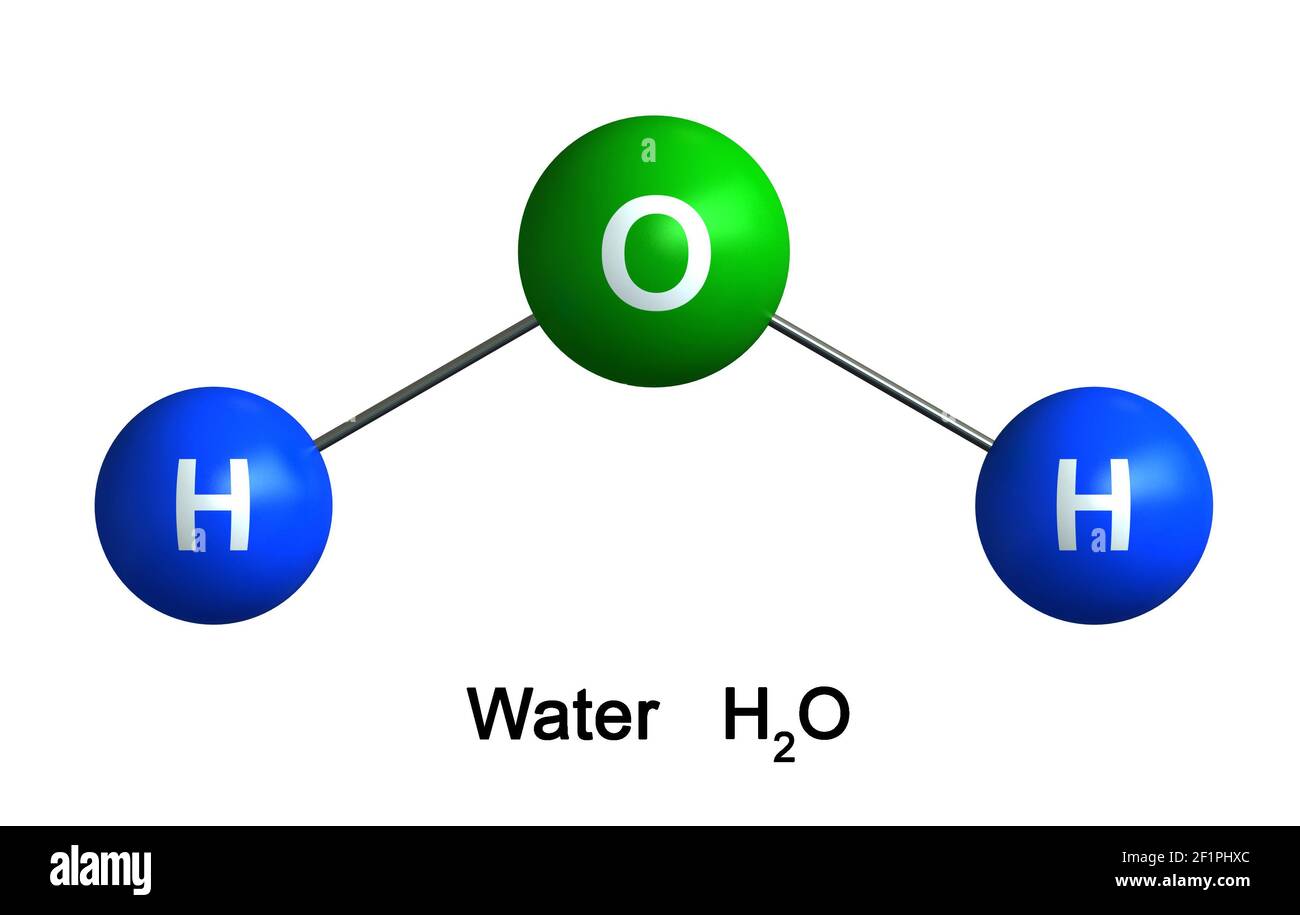
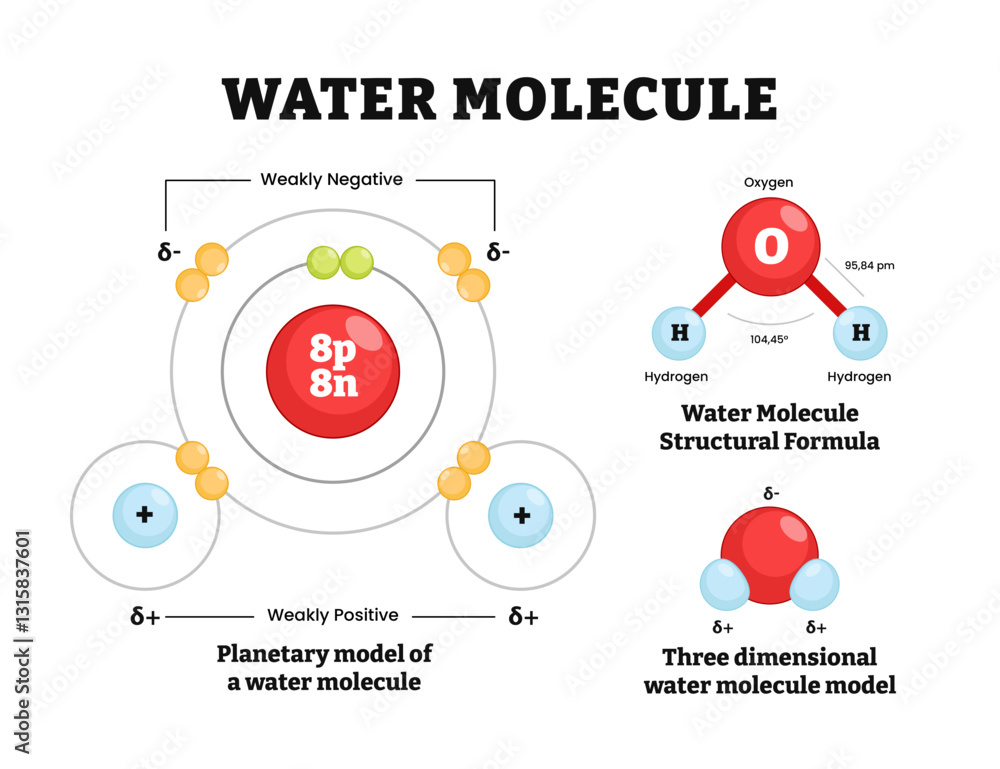
The Molecular Structure Of Water comes down to a single oxygen atom bonded to two hydrogen atoms, but the geometry matters more than you might expect. The O-H bond length sits at about 0.957 angstroms, and the angle between them is roughly 104.5 degrees. That bent shape isn't arbitrary—it comes directly from the sp3 hybridization of the oxygen atom's valence shell, which carries two lone pairs pushing against the bonding pairs. That bond angle deviates from the ideal tetrahedral angle of 109.5 degrees because the lone pairs occupy more spatial volume than bonding pairs, compressing the H-O-H angle. This asymmetry gives water a permanent dipole moment of about 1.85 debyes. The partial negative charge sits on the oxygen, partial positive on each hydrogen. That charge separation is why water does almost everything it does—dissolving salts, forming hydrogen bond networks, staying liquid at room temperature instead of being a gas like HS. The practical consequence nobody stresses enough is that water doesn't have a fixed, static structure in the liquid phase. Each molecule forms roughly 3.4 hydrogen bonds on average at room temperature, but those bonds break and reform on a picosecond timescale. The instantaneous arrangement looks tetrahedral some of the time and distorted the rest of the time. Modeling this accurately in a simulation is harder than it sounds.
I spent weeks debugging a molecular dynamics run where the density of liquid water kept coming out 2 percent too high. Standard TIP3P force field, nothing fancy. The issue wasn't the code or the integration step. It was the box size. When you're running water simulations with periodic boundary conditions and the box drops below about 2 nanometers on a side, the artificial self-correlation of molecules through their own periodic images starts shifting the radial distribution function g(r) noticeably. The first peak gets sharper, the density shifts up. I increased the box to 4 nanometers and the density correction dropped to within 0.3 percent of the experimental 0.997 g/cm³. The simulation runtime went from a few hours to roughly twelve hours on the same hardware, so there's a real tradeoff here depending on what you're trying to measure. There's a common misconception that the hydrogen bonds in water are directional and rigid like covalent bonds. They're not. A typical hydrogen bond in liquid water has an energy of about 20 kilojoules per mole and allows angular fluctuations of roughly 30 degrees before it breaks entirely. That flexibility is why ice has a lower density than liquid water—the tetrahedral lattice in ice locks the bonds into a more open arrangement. When ice melts, some of those angular constraints relax and molecules can pack closer. This density inversion only works because of the specific Molecular Structure Of Water geometry. Most substances get denser when they melt. Another thing that trips people up is the isotope effect. Heavy water (DO) has a bond angle of about 104.48 degrees and a slightly shorter O-D bond at 0.957 angstroms, essentially the same as HO. The differences show up in bulk properties instead—DO has a higher boiling point at 101.4°C compared to 100°C, and a density of 1.105 g/cm³ at 25°C. The quantum mechanical reason is zero-point vibrational energy. Deuterium is twice as heavy as protium, so its ground-state vibrational amplitude is lower. That subtly stiffens the hydrogen bond network and shifts the thermodynamic properties. If you're doing isotope-labeling experiments or comparing simulation results to experimental data, this difference matters even though the structural parameters look identical on paper.
When you move beyond classical force fields and try ab initio molecular dynamics using density functional theory, you run into the self-interaction error problem. Standard functionals like PBE underestimate the hydrogen bond strength by about 15 to 20 percent, which means the simulated water is too diffuse and the diffusion coefficient comes out roughly twice the experimental value of 2.3 × 10 m²/s. Adding a dispersion correction like DFT-D3 brings the structure into line but adds computational overhead. For most practical purposes, a well-parameterized empirical potential like TIP4P/2005 gives you radial distribution functions within 5 percent of experimental neutron scattering data at a fraction of the cost. The TIP4P/2005 model places a virtual site along the bisector of the H-O-H angle to better represent the lone pair electron density, which is why it handles the liquid structure better than the older TIP3P model despite both being four-site potentials. X-ray and neutron diffraction remain the primary experimental methods for determining the Molecular Structure Of Water in the liquid state. X-ray scattering gives you information weighted by the atomic form factors, so oxygen dominates the signal. Neutron scattering depends on nuclear scattering lengths, which vary irregularly across the periodic table—hydrogen has a negative scattering length while deuterium is strongly positive. That's why isotopic substitution experiments (swapping H for D) are standard practice in liquid water structure studies. They let you isolate the H-O and H-H correlation functions that X-ray data alone can't resolve. The second coordination shell sits at about 4.5 angstroms from the central oxygen and contains roughly five molecules. That shell is important because it mediates the long-range correlations that give water its unusually high bulk modulus and its anomalous heat capacity. Some researchers have argued that water might harbor a second critical point at deeply supercooled temperatures near -45°C, which would explain the growing correlation length seen in simulations. Whether this is real physics or a force-field artifact is still debated, and the experimental access to that temperature regime is extremely limited because water crystallizes before it gets there.
Get the Full Details

If you're working with this in a lab or a simulation, the main thing to keep straight is that the Molecular Structure Of Water you measure depends on the method and the conditions. Gas-phase clusters, liquid water, ice Ih, and supercritical water all show different average bond angles and coordination numbers. There's no single answer that applies everywhere. The bent geometry and the dipole are the constants. Everything else comes from how those molecules arrange themselves under whatever constraints you're putting them under.