Molecular Orbital Diagram of O2 Explained
Most textbooks show the MO diagram for oxygen and then hand-wave through why O2 is paramagnetic. Here's how it actually works in practice and where people consistently get tripped up.Mot Diagram Of O2
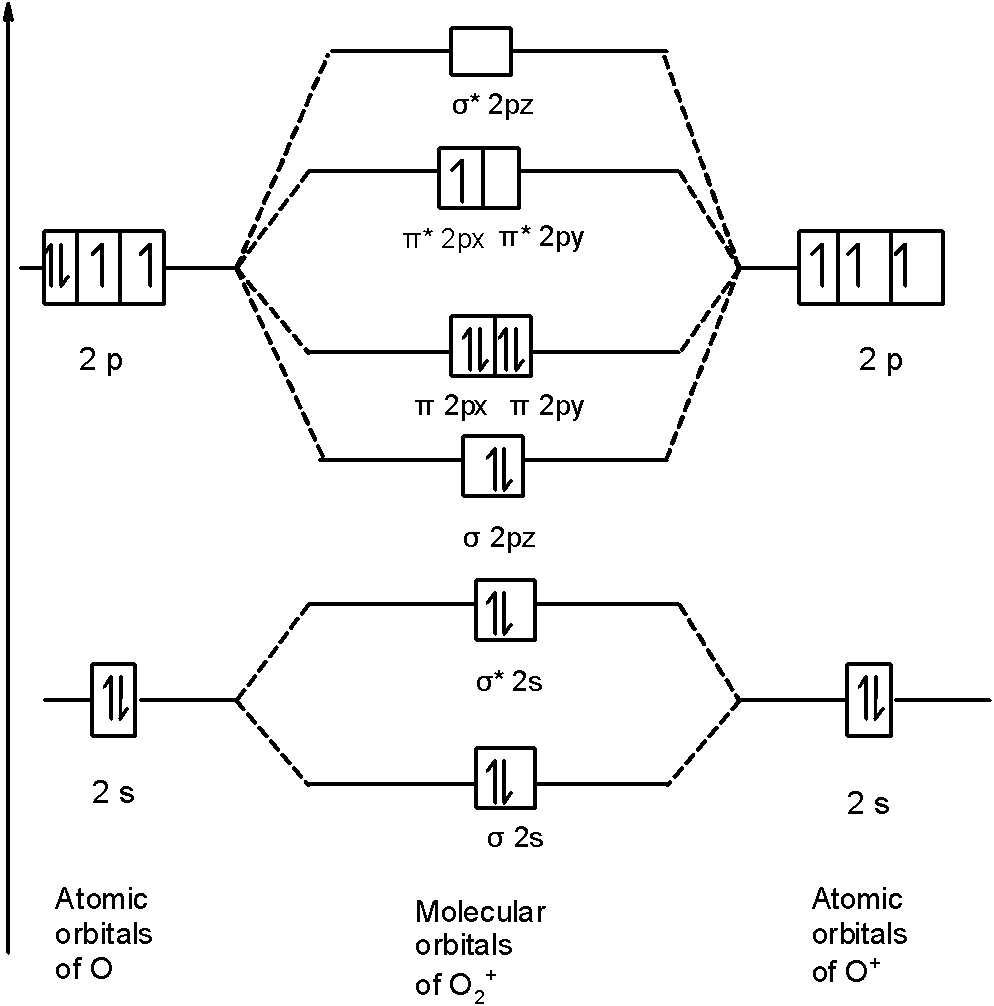
The diagram starts with two oxygen atoms, each contributing six valence electrons. That's twelve total. You fill the molecular orbitals in this order: (2s), *(2s), (2pz), (2px) and (2py) — these two are degenerate — then *(2px) and *(2py), which are also degenerate, and finally *(2pz). The first four orbitals are straightforward. (2s) holds two electrons, *(2s) holds two, (2pz) holds two. Then you put four electrons into the two bonding orbitals. That accounts for ten electrons. The last two go into the * antibonding orbitals. Here's where it matters: according to Hund's Rule, those two electrons occupy separate * orbitals with parallel spins rather than pairing up in one. That gives you two unpaired electrons. O2 is a triplet diradical. This is why liquid oxygen sticks between the poles of a magnet. The bond order calculation confirms what we already know experimentally. You have eight bonding electrons and four antibonding electrons. Eight minus four, divided by two, equals two. Double bond. Matches the Lewis structure, though Lewis structures completely fail to predict the paramagnetism.A common mistake is drawing the (2pz) orbital below the (2p) orbitals the way you would for N2. For oxygen and fluorine, the s-p mixing effect is negligible, so the (2pz) drops below the (2p) set. Students who memorize the nitrogen diagram and apply it to oxygen will get the ordering wrong and end up with no unpaired electrons. That's the most common error I see in first-year exams.
I encountered a specific issue last semester when a student was trying to calculate the magnetic moment using the formula = (n(n+2)) Bohr magnetons, where n is the number of unpaired electrons. They plugged in n=0 because they had drawn the diagram with the wrong orbital ordering. When we corrected the diagram, n=2, giving = 8 2.83 BM, which matches the experimental value almost exactly. The disagreement usually comes from forgetting that O2 has that slight deviation due to spin-orbit coupling, but for most purposes 2.83 is accurate enough.