Working With Nucleotides In Practice
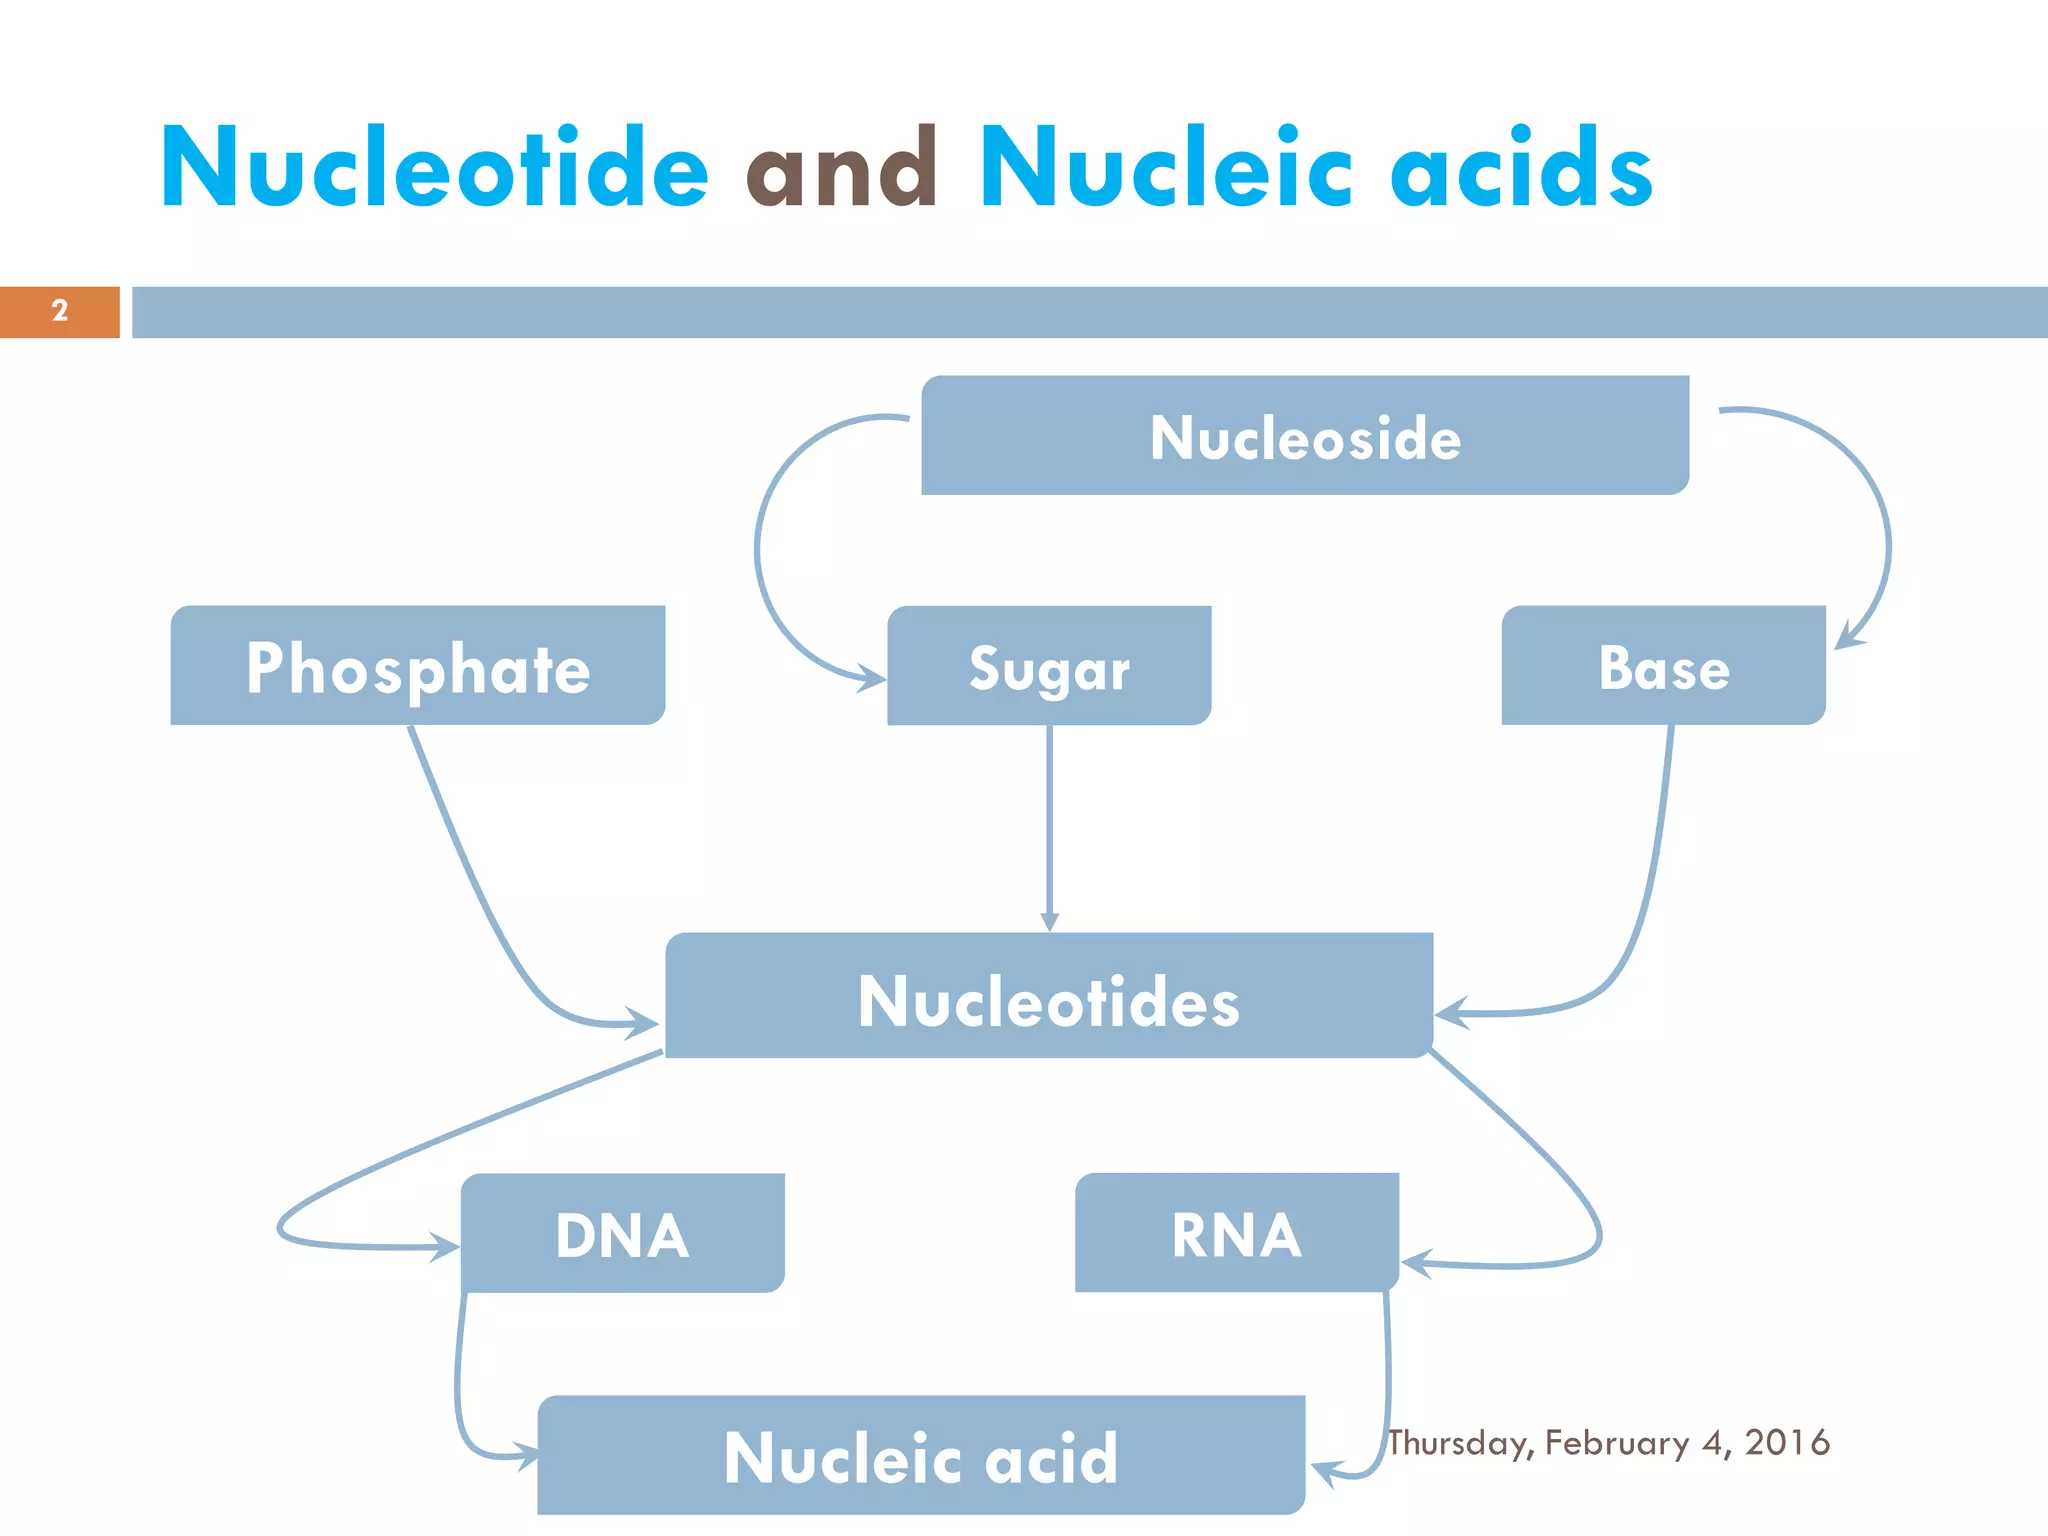
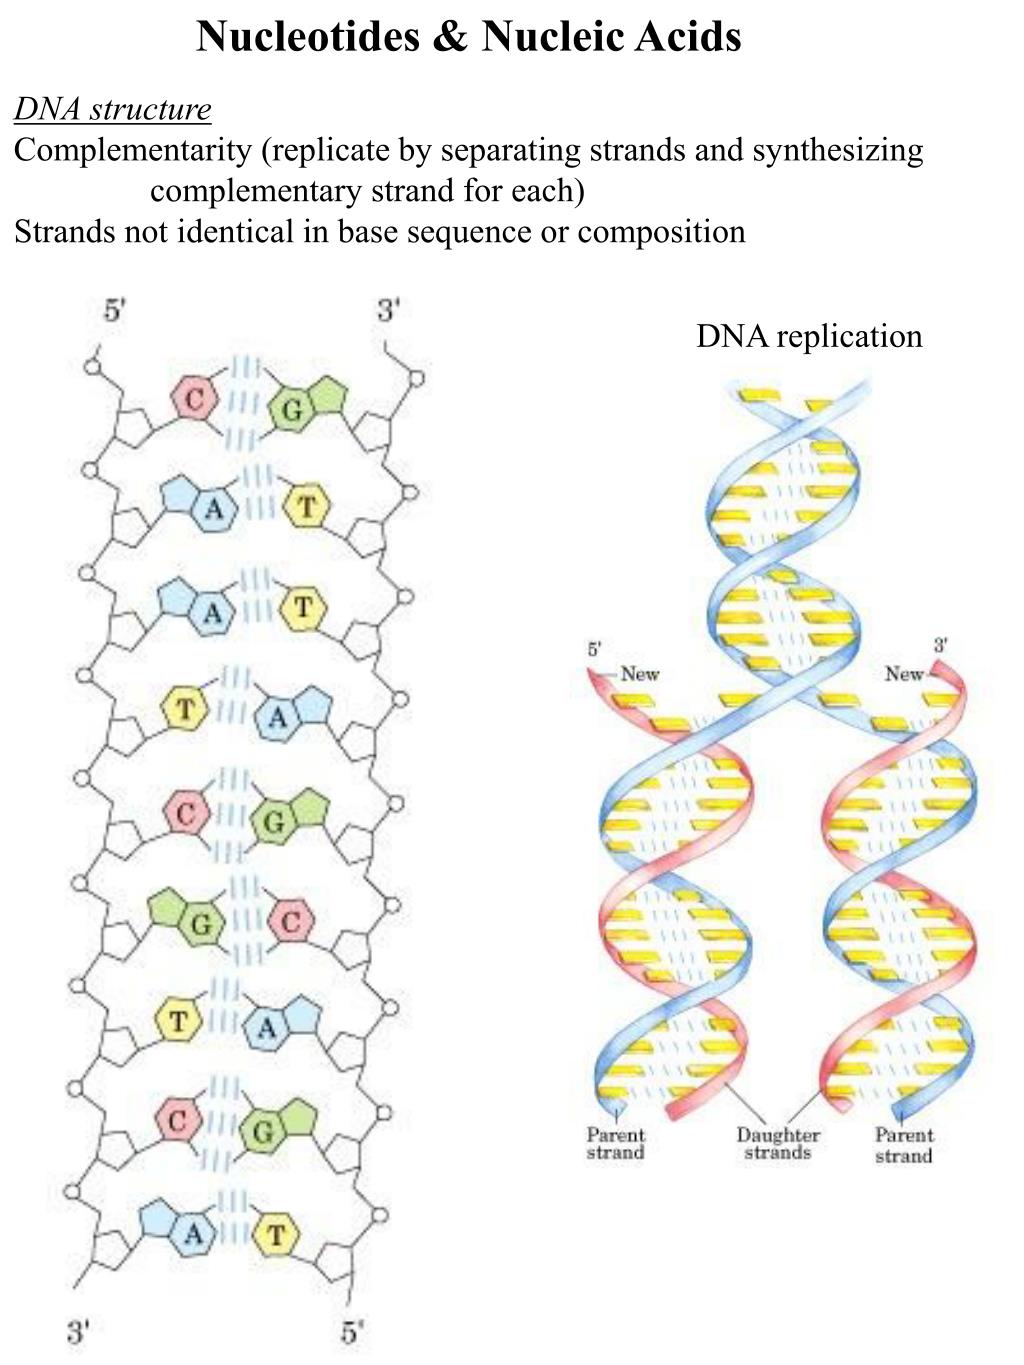
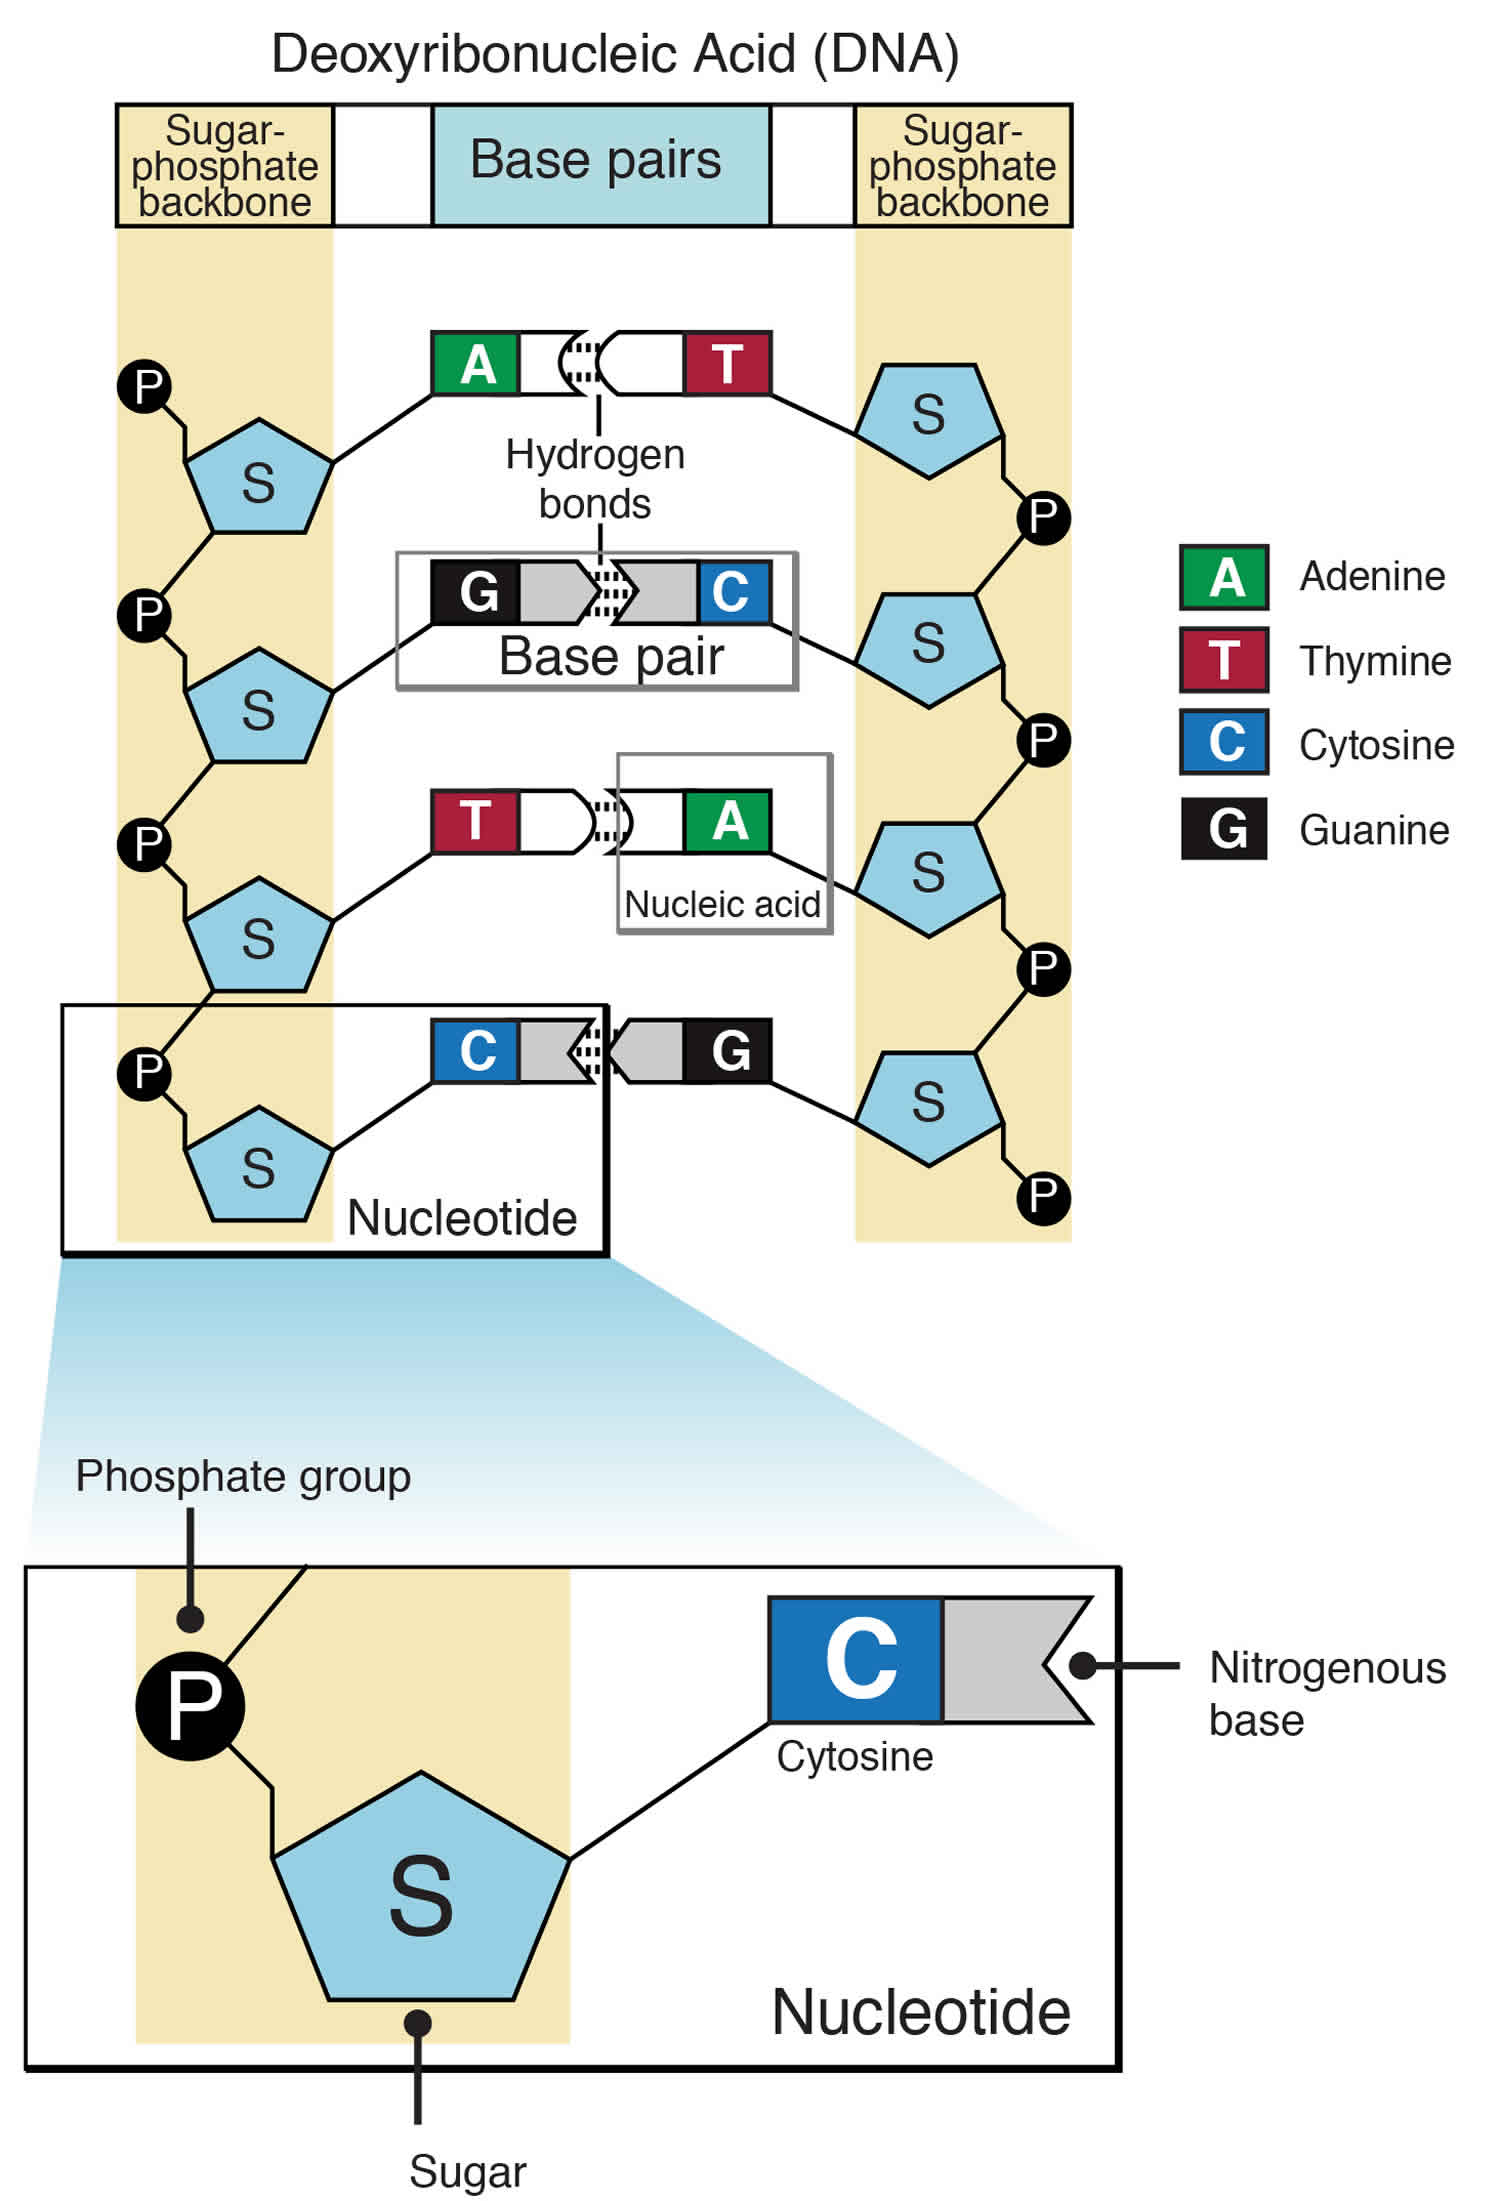
Most people coming into this field learn the basics and then move on without really understanding what happens when you try to use them in a lab setting. The gap between textbook nucleotide And Nucleic Acid explanations and actual bench work is where most mistakes happen. I spent years troubleshooting PCR failures before I realized the issue wasn't my primer design or thermal cycler it was the nucleotide quality and concentration in my master mixes. A nucleotide consists of three components: a nitrogenous base, a five-carbon sugar, and at least one phosphate group. When these link together through phosphodiester bonds they form nucleic acids like DNA and RNA. That is the simple version. The complicated version involves dNTPs versus rNTPs, modified bases, and the fact that not all nucleotides behave the same way under different buffer conditions. I learned this the hard way during a project where I was running long-range PCR amplifications. My products kept showing smears on gels despite perfect primer specs and clean template DNA. After weeks of swapping reagents I isolated the problem to degraded dNTPs in an old aliquot that had been freeze-thawed too many times. Nucleotide degradation creates misincorporation events and reduces polymerase processivity. I switched to fresh aliquots and the smearing stopped immediately.
Setting Up Your Nucleotide Solutions Correctly
The most common mistake I see is people making working stocks that are too concentrated and then storing them at -20°C for months. Each freeze-thaw cycle degrades roughly 5 to 10 percent of your nucleotide supply. Make 100mM stocks from solid nucleotides using the appropriate buffer, aliquot into small volumes, and never go above 10 rounds of freeze-thaw. For routine PCR a final concentration of 200µM each dNTP works for most applications. Higher concentrations can actually inhibit Taq polymerase and increase misincorporation rates. Some protocols call for asymmetric PCR with imbalanced dNTP concentrations but that requires optimization and is not something to attempt without understanding your polymerase's tolerance.
Common Pitfalls Nobody Talks About
Nucleotide purity matters more than concentration. Cheap dNTP mixtures often contain free phosphate and residual solvents from the manufacturing process. These contaminants chelate magnesium ions in your reaction buffer. A typical MgCl2 concentration of 1.5mM can drop to effectively 0.8mM if your dNTPs are impure. That single issue explains so many inconsistent amplification results. Another thing people overlook is the pH of your nucleotide solutions. dNTPs should be dissolved in Tris buffer adjusted to around pH 7.0 to 7.5. If the pH drifts too low the nucleotides precipitate out of solution. Too high and you risk deamination of the adenine bases. I once ran a sequencing reaction where the read quality dropped sharply in the middle of the template and traced it back to a slightly acidic dNTP stock that had lost buffering capacity over time. There are situations where nucleotide analogs become necessary. Modified bases like bromodeoxyuridine or fluorescently labeled terminators serve specific purposes but they change the kinetics of polymerization. A regular polymerase will stall or terminate differently with these analogs. If you are doing primer extension assays or next-generation sequencing library prep you need to verify your polymerase compatibility with any modified nucleotides you introduce.
Get the Full Details
When Standard Approaches Fail
Some applications simply cannot use standard dNTPs. Long amplicon sequencing, single-molecule real-time sequencing, and certain mutagenesis protocols require specialized nucleotide mixtures. In those cases the cost difference between generic and validated reagents becomes irrelevant because a failed reaction costs more than the reagent upgrade. For routine cloning and Sanger sequencing I recommend sticking with well-established commercial dNTP sets from vendors who provide lot-specific purity data. The extra cost over unverified suppliers is usually less than 50 dollars per order and it saves significant time on troubleshooting. If you are running high-throughput workflows the yield consistency from validated lots becomes economically significant over a year. Nucleotide And Nucleic Acid chemistry is straightforward until you need it to be precise. The details around storage, purity, and concentration are what separate reproducible results from frustrating variability. Focus on those factors first before you blame your protocols or equipment.