
Working With Of State Chemistry in Practice
When I first started dealing with phase behavior in industrial systems, most people treated it like textbook material. It isn't. The gap between the Clausius-Clapeyron equation and what actually happens when you're running a 40-bar distillation column at midnight is substantial. That's where Of State Chemistry lives for anyone who has to make things work outside the lab. The core idea is straightforward enough: matter changes properties when you move between solid, liquid, and gas phases, and those transitions aren't always clean. Temperature, pressure, and composition interact in ways that can wreck your schedule if you don't account for them early. I've watched teams lose three weeks because they assumed a binary mixture would behave ideally across the entire operating window. It didn't. They ended up recalibrating their pumps, replacing seals, and rewriting their run sheet.
What Of State Chemistry Actually Means for Your Work
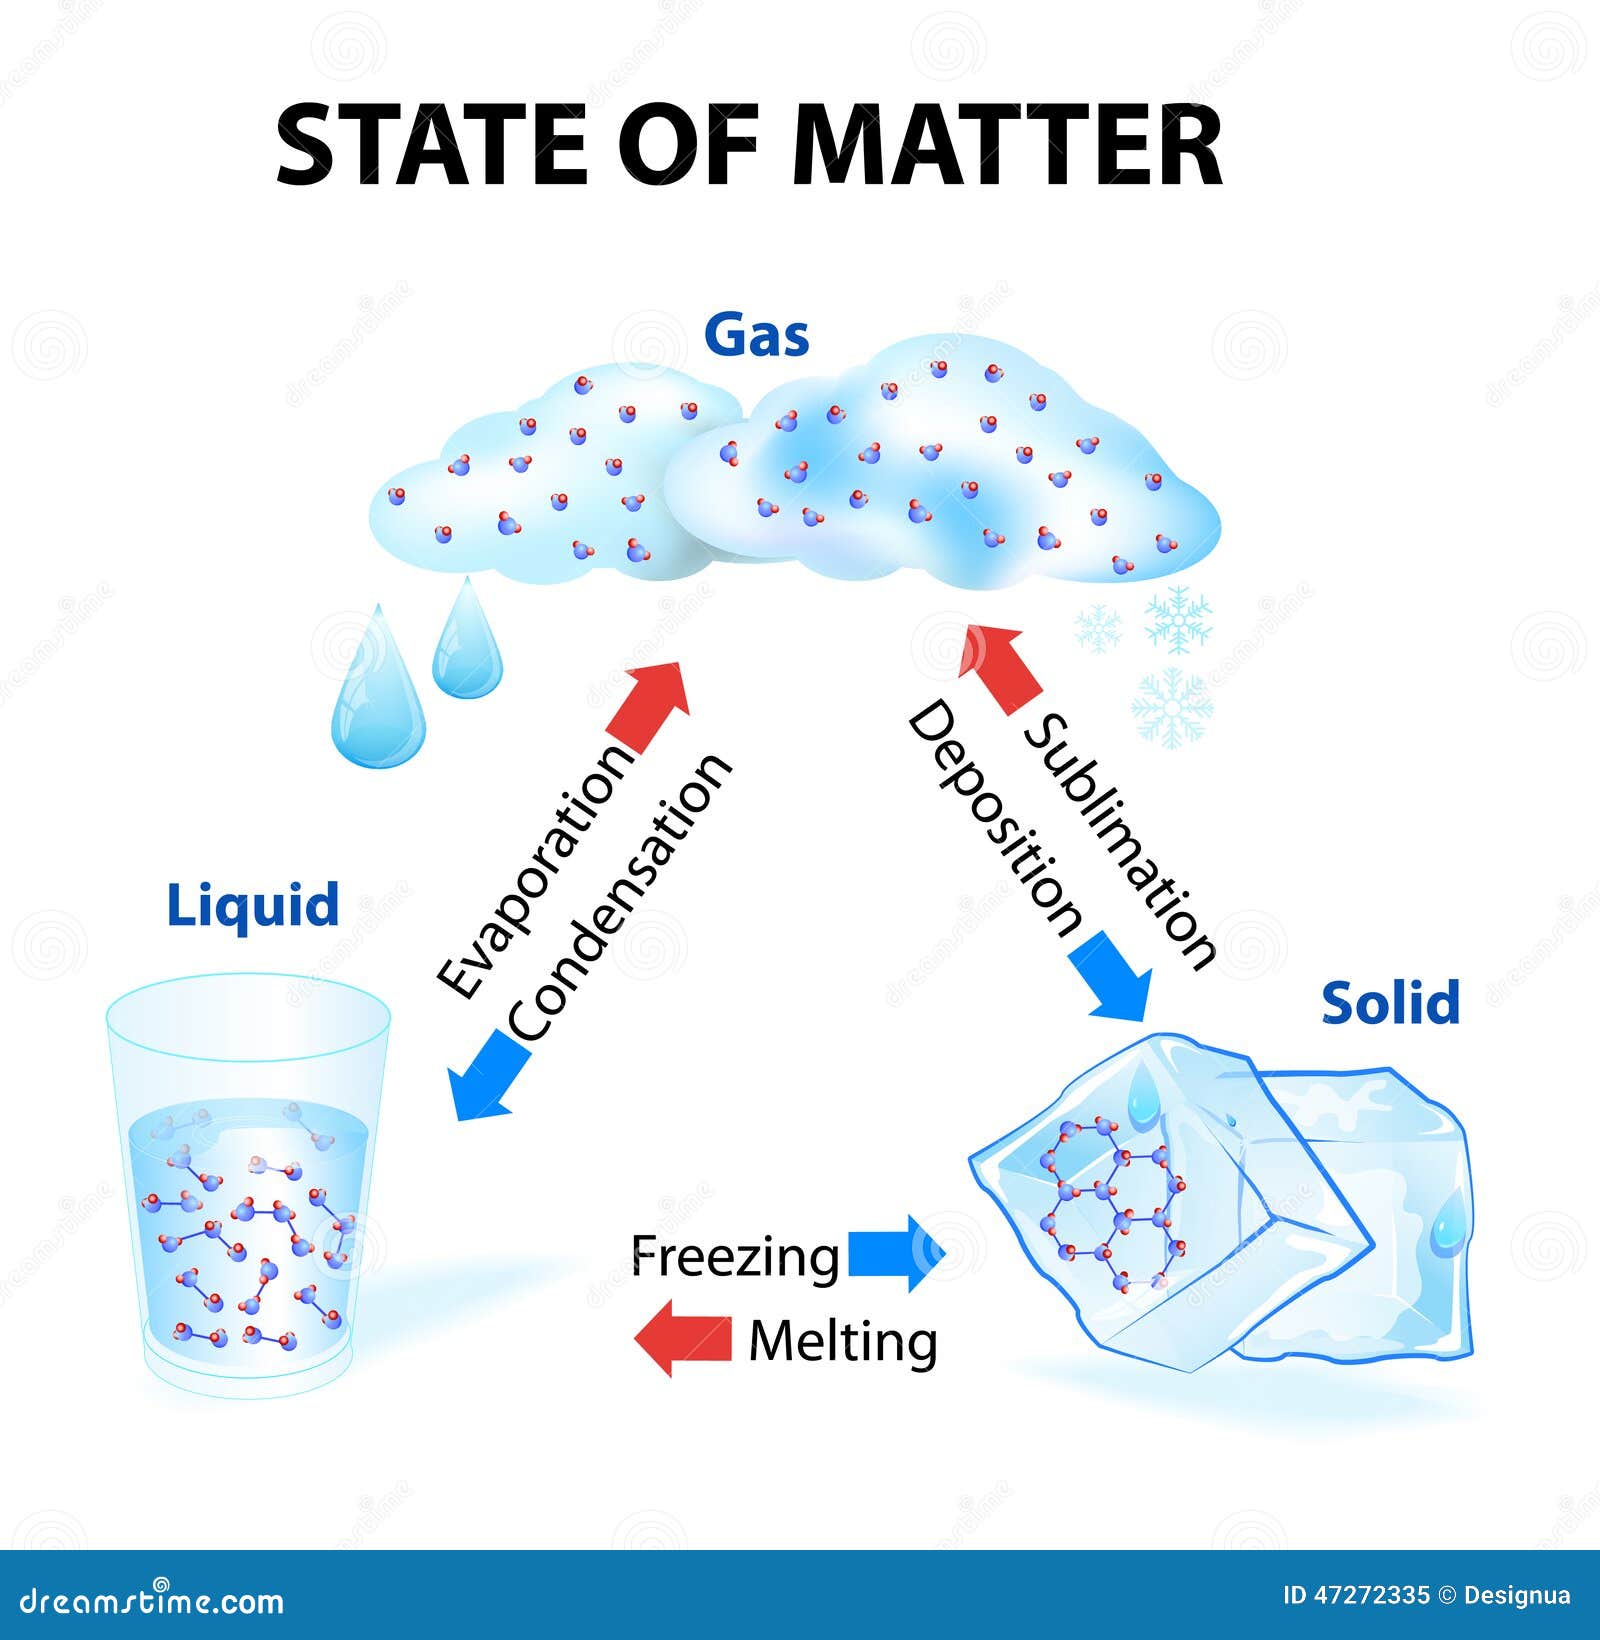
Of State Chemistry isn't a single technique. It's the practical application of understanding how substances behave across different phases and using that knowledge to control processes. This covers things like predicting when a solvent will precipitate out of solution, calculating the pressure needed to keep CO2 supercritical, or figuring out why your reaction mixture is flashing to gas at a point you thought was safely liquid. The most important tool you'll reach for is the phase diagram. Not the idealized version from your undergraduate text, but the real one built from experimental data or validated simulation. Aspen Plus and HYSYS are standard in the industry, but they're only as good as the parameter sets you feed them. I learned that the hard way when a colleague ran a crude oil assay through the default NBF (National Bureau of Standards) fractions and got boiling point predictions that were off by nearly 40 degrees. We rebuilt the heavy end using actual residue analysis from the supplier, and the model clicked into place.
How to Approach It When You're Starting
Start by mapping your system. Identify every component, then figure out which ones matter for your operating range. Water usually matters everywhere. Organic solvents matter in specific windows. Trace impurities can dominate behavior in narrow ranges and disappear in others. You need to know which is which before you design anything. Next, get the P-T-composition data you need. If you can't find it in the literature or a database like DIPPR or NIST Chemistry WebBook, you're going to have to measure it. There's no shortcut. A student I mentored once tried to model a glycol-water-ethanol system using only pure-component parameters and wondered why his azeotrope predictions were garbage. The excess Gibbs energy interactions between those three components are significant and well-documented. He just didn't look them up. After that, build your model. Use the right activity coefficient method for your system. NRTL works for many liquid-liquid equilibria problems. UNIQUAC handles polymer systems better. Peng-Robinson is standard for vapor-liquid work at moderate pressures. Each has failure modes. The Wilson equation breaks down for partially miscible systems. Margules is too simplistic for anything beyond rough estimates. Pick carefully.
Get the Full Details
Finally, validate against something real. A literature boiling point. A pilot plant reading. A single data point from your own lab. If your model can't reproduce one thing you know is true, don't trust it for anything else.
A Specific Problem I Ran Into
About five years ago, I was troubleshooting a crystallization step for a pharmaceutical intermediate. The process called for cooling a methanol-water solution from 60 degrees Celsius down to near zero to precipitate the product. Everything looked fine on paper. The solubility curve was well documented. The cooling rate was conservative. Except the material wasn't coming out of solution. I spent two days digging through it before I realized what was happening. The solution had become supersaturated, yes, but the nucleation barrier was too high at those temperatures without a seed crystal. The compound was sitting there in stable suspension, completely metastable, refusing to crash out. We added a small charge of pre-ground product as seed, and the yield hit 94 percent on the next batch. Without that, we were looking at either waiting days for spontaneous nucleation or forcing the temperature lower and risking solvent incorporation into the crystal lattice. This is the kind of thing that doesn't show up in phase diagrams. It shows up when you're actually running the process and staring at a reactor full of clear liquid wondering where your product went.
Common Mistakes That Cost Time
One of the biggest is ignoring non-ideal behavior in mixtures. People assume Raoult's law is close enough until it isn't, usually at the worst possible moment. Acetone-chloroform is a classic example of positive deviation from ideality with a maximum-boiling azeotrope. If you design a separation column assuming ideal behavior, you'll never get the purity you specified. Another is treating pressure as secondary. At low pressures, it often is. At high pressures, it dominates. Supercritical fluids are the obvious case, but even moderate pressurization can shift boiling points enough to matter. I once saw a team run a reflux condenser at atmospheric pressure when their system was operating at 3 bar gauge, wondering why their overhead compressor was cavitating. They hadn't accounted for the temperature-pressure relationship in their condenser duty calculation. A third mistake is relying on old data without checking it against current standards. Many of the activity coefficient parameters floating around in older textbooks were derived from experiments done with equipment that wouldn't pass inspection today. Temperature control, pressure measurement, equilibrium verification — the uncertainty bands were wider than people realized. If you're working with regulatory compliance or tight specifications, verify your parameters against newer sources.

Where It Falls Short
Of State Chemistry has real limitations. It works well for systems with known thermodynamic parameters and moderate conditions. It gets shaky when you're dealing with reactive mixtures, polymer solutions, or conditions near critical points where property gradients become extreme. It also assumes equilibrium, which is rarely true in fast industrial processes. A flash drum operates far from equilibrium in practice, and models that don't account for mass transfer limitations will overpredict separation efficiency. For complex reactive systems, you may need to combine thermodynamic modeling with kinetic analysis. For polymer systems, equations of state like PC-SAFT are more appropriate than traditional activity coefficient models. For systems near criticality, you need multiphase flow simulations that capture the hydrodynamics, not just the equilibrium thermodynamics.
The Practical Value of Of State Chemistry
Despite its limitations, Of State Chemistry is the foundation for almost everything that happens in process design and optimization. Without it, you're guessing. With it, you can predict, troubleshoot, and scale with confidence. The skill isn't in memorizing equations. It's in knowing which equations apply, when they break, and what to do when they do. The people who get good at this aren't the ones who read the most textbooks. They're the ones who've seen enough failures to know what to watch for. A phase envelope that closes earlier than expected. An azeotrope that shifts with pressure. A solid form that polymorphs under the wrong conditions. These are the things that teach you more than any perfect model ever will. If you want to start practicing, pick a simple system you understand and break it. Run a calculation where the answer is wrong on purpose. Then fix it. That's how the intuition builds.


