Getting the Protection Right the First Time
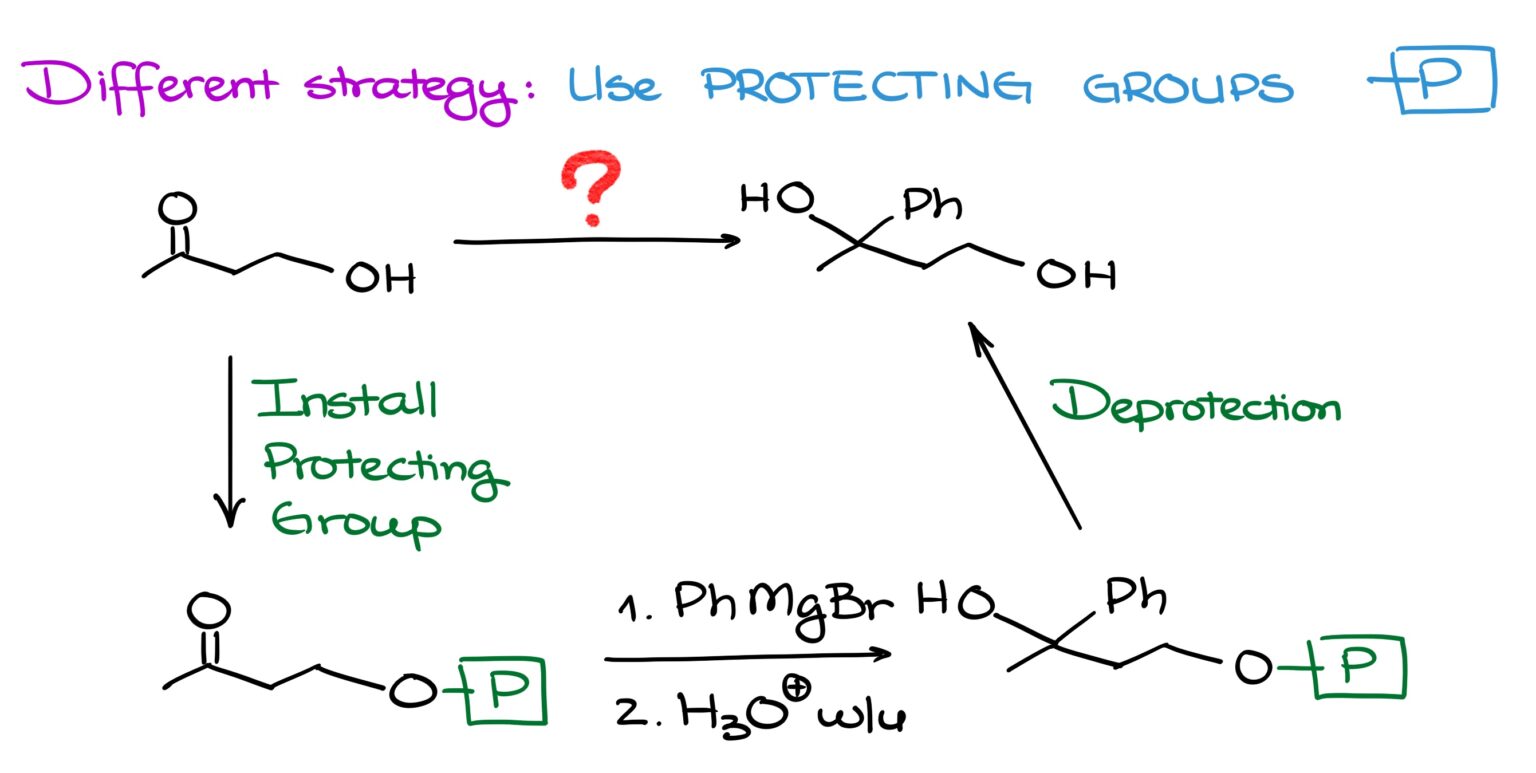
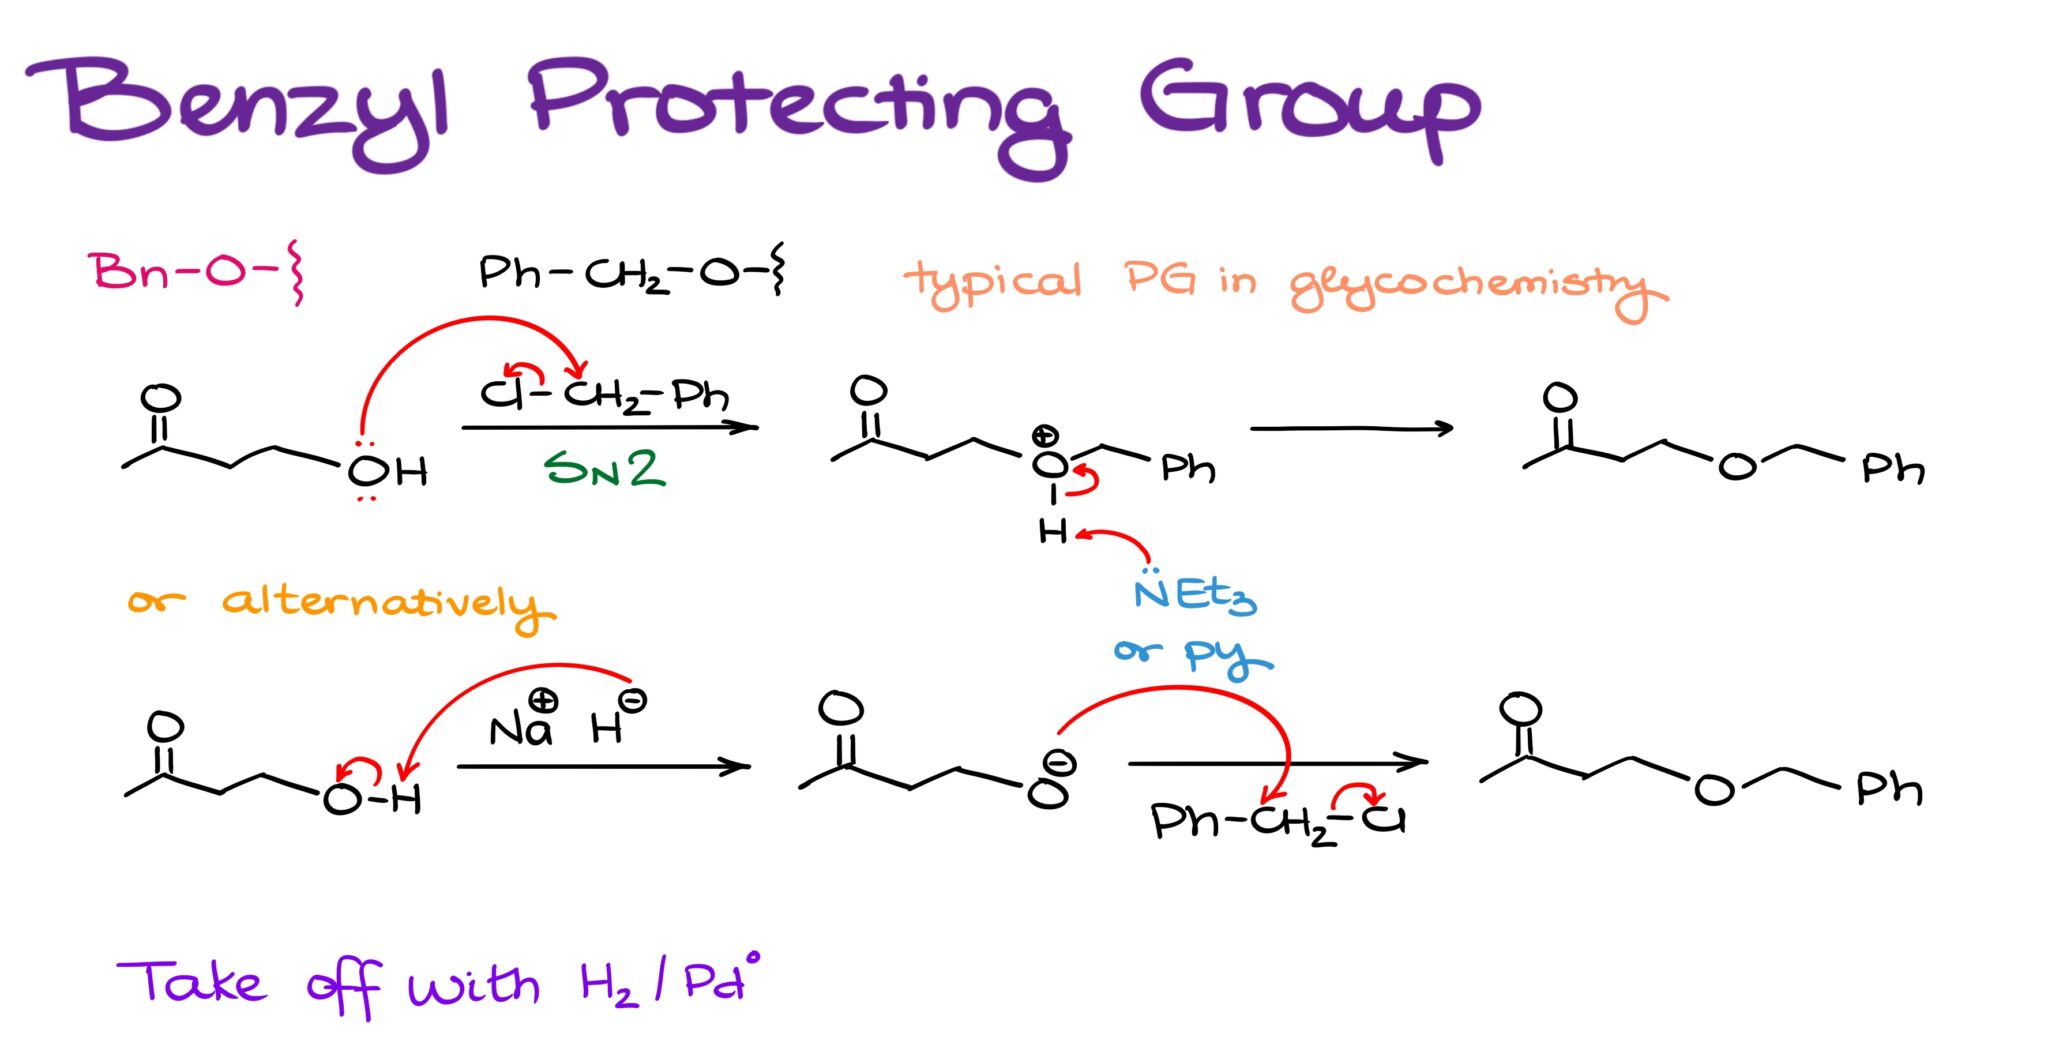
Protecting groups are one of those things you learn the hard way. You think you understand them in lecture. Then you are standing over a flask at 2 AM watching your reaction turn into tar because you did not account for chemoselectivity. I have spent years working through exactly where these reagents fail and where they work. This is what actually matters. The core idea is simple enough. You have a molecule with two reactive sites and you need to transform one without touching the other. So you temporarily mask the site you do not want to react. The protecting group has to go on cleanly, survive whatever conditions your synthesis throws at it, and come off without damaging the rest of the molecule. That last part is where most people get burned. I still remember a project where I was building a multi-step sequence on a intermediate with both a free hydroxyl and a secondary amine. I needed to acylate the amine selectively. Everyone defaults to Fmoc or Boc on the nitrogen, but those are designed for removal under basic or acidic conditions respectively. My downstream chemistry required a strong base. So I went with a carbamate strategy using Cbz, which tolerates base but comes off with hydrogenation. The problem was my substrate had a benzyl ether elsewhere that also wanted to get hydrogenolyzed. I ended up switching to a para-methoxybenzyl (PMB) ether for that position because it survives catalytic hydrogenation but cleaves selectively with DDQ. That adjustment cost me three extra chromatography runs and a full day of troubleshooting, but it saved the whole sequence. If I had known earlier, I could have planned around it.
How to Actually Choose a Protecting Group
Most guides will just list protecting groups alphabetically. That is useless when you are designing a route. The decision tree is more like this: first identify every functional group in your molecule that could interfere. Then map your synthetic steps and note the conditions each one requires. After that, find the overlap — which protecting groups survive those conditions and can be removed selectively. Strong acid conditions? Avoid tert-butyl ethers and acetals. Strong base? Acetates and silyl ethers might not hold up. Oxidizing environments? Thioethers and some sulfonyl groups are out. Reducing conditions like hydrogenation with palladium? Benzyl-based protections will fall off. You need to know this before you put anything on. One thing beginners consistently miss is the order of removal. If your molecule has two different protecting groups, you need to be able to remove them independently without affecting the other. That is called orthogonal protection and it is not optional for any multi-step synthesis. The classic orthogonal pair is a tert-butyldimethylsilyl (TBS) ether on oxygen and an Fmoc carbamate on nitrogen. TBS comes off with fluoride. Fmoc comes off with mild base like piperidine. They do not interfere with each other at all.
The Silyl Ether Problem Nobody Talks About
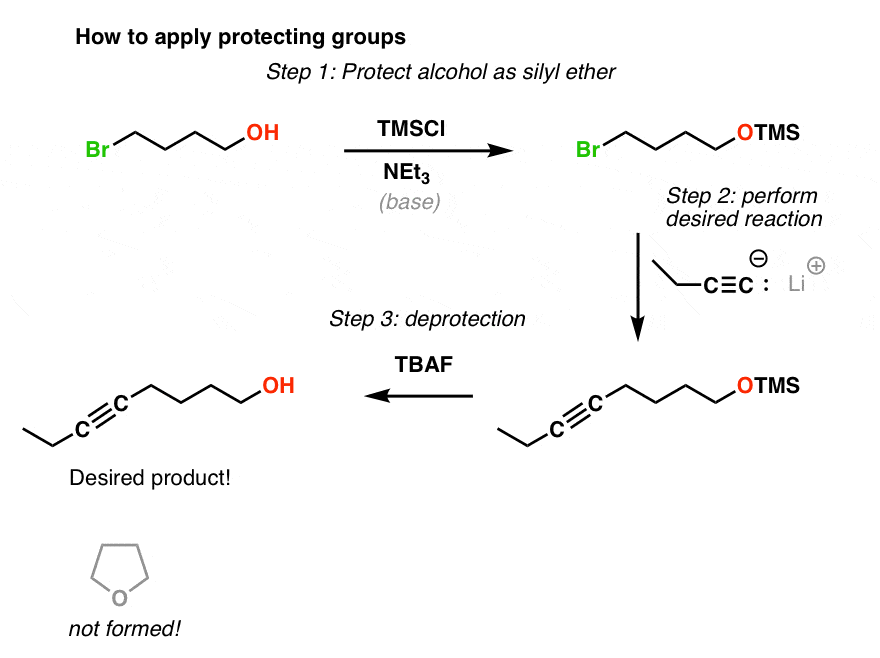
TBS, TMS, TBDPS, TES — these are the most common silyl protecting groups for alcohols. They are convenient, they go on fast, and they come off with fluoride sources. But there is a catch. Silyl ethers are sensitive to nucleophiles and to certain basic conditions that are not obviously basic. I once ran a Mitsunobu reaction on a TBS-protected alcohol and watched the protecting group partially cleave. DIAD and triphenylphosphine in THF at room temperature is apparently harsh enough to nibble at TBS over several hours. Switching to TBDPS fixed it because the bulkier diphenylisopropyl group is significantly more robust. That is the tradeoff. More stability often means harder removal later. Acetals and ketals are another category that deserves more caution. Dimethoxymethane with p-toluenesulfonic acid gives you a MOM ether. 1,3-propanediol gives you a cyclic acetal. These are fine for acid-sensitive substrates, but they are unstable in aqueous acid. If your workup involves anything even mildly acidic and aqueous, you could lose your protection during the extraction phase. I have seen students report zero yield and spend two days figuring out the protecting group fell off during the brine wash. Use a non-aqueous workup or switch to a more robust group if your chemistry demands it.
Get the Full Details
When Standard Deprotection Fails
There are times when the textbook deprotection conditions just do not work. Your substrate might have another functional group that reacts with the standard reagent. Or the protecting group might be more tightly bound than expected due to steric effects. In those cases you need alternatives, and knowing them saves you from starting over. For example, removing a Boc group typically uses trifluoroacetic acid in dichloromethane. But if your molecule has a base-sensitive ester somewhere else, the TFA will not bother it, so that is fine. If instead you have an acid-sensitive acetal, TFA will destroy it. The workaround is to use hydrogen chloride in dioxane or even milder acids like camphorsulfonic acid. It takes longer but it is selective enough. I once had a substrate with a Boc group and a pendant orthoester. Standard TFA deprotection collapsed the orthoester into an aldehyde. Switching to HCl in dioxane at 0 degrees Celsius gave clean deprotection in about 45 minutes with no side reactions. The orthoester stayed intact. Fluoride removal of silyl groups usually uses TBAF in THF. That is the default. But TBAF is somewhat basic and it can cause elimination on secondary or benzylic positions. If you are dealing with a substrate that has a beta-hydrogen and a good leaving group nearby, TBAF might trigger an elimination side reaction. The fix is to use a milder fluoride source like CsF in DMF or acetonitrile. It is slower, sometimes taking 2 to 4 hours instead of 30 minutes, but it avoids the basicity problem entirely. I use CsF now as my default for anything that looks remotely elimination-prone.
Pitfalls That Waste Days
The most common mistake is installing the protecting group and then realizing too late that the deprotection conditions wreck another part of the molecule. This is especially painful with boronic esters and other organoboron intermediates. If you protect an alcohol as a TBS ether and then need to do a Suzuki coupling later, the fluoride from deprotection will also cleave the boronic ester. You cannot deprotect after the coupling unless you regenerate the boron, which is messy. The solution is to either use a protecting group that comes off without fluoride, like an acetate removed with base, or to install the boron after deprotection rather than before. Another issue is incomplete protection. You add the protecting group reagent, run the reaction, and assume it went to completion because the starting material peak is gone on TLC. But sometimes you have half-protected material co-eluting or running just slightly differently. If you carry that into the next step, you end up with a mixture of protected and unprotected starting material reacting at different rates. I always run a quick derivatization check, like acetylation with acetic anhydride and DMAP, to confirm all hydroxyls are masked. Unprotected hydroxyls will acetylate rapidly and shift the spot on TLC. It takes five minutes and saves you from chasing impurities later.
A Quick Reference That Actually Works
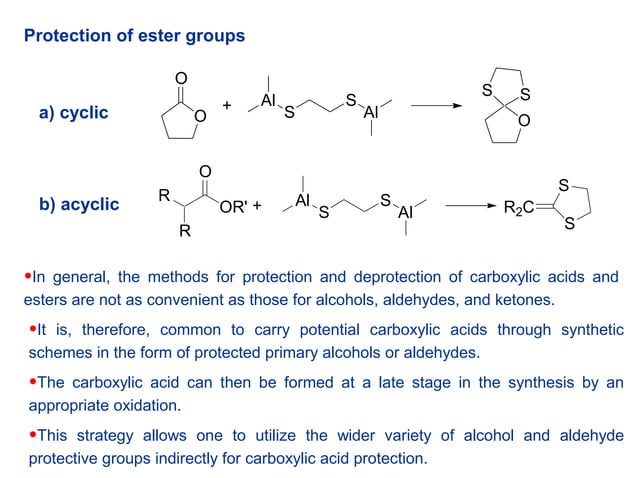
Hydroxyls: TBS for general use, TBDPS for base stability, MOM for acid stability but fluoride-free deprotection is impossible, acetate for easy base removal but not robust to many conditions. Amines: Boc for acid removal, Fmoc for base removal, Cbz for hydrogenolytic removal. All three are standard. Pick based on your downstream chemistry, not habit. Aldehydes and ketones: Acetals and ketals for acid stability, dithianes if you need something even more robust but removal requires heavy metals like mercury or silver which complicates purification.

Carboxylic acids: Methyl esters, ethyl esters, tert-butyl esters, and benzyl esters each require different deprotection. Methyl and ethyl need strong base hydrolysis. Tert-butyl comes off with acid. Benzyl needs hydrogenation. Choose based on what your subsequent steps can tolerate.
Bottom Line
Protecting groups are not a minor detail. They are central to almost every multi-step organic synthesis. The key is to plan them backward from your final step. Know how you will remove each one before you put it on. Check that the removal conditions do not conflict with other protections or functional groups in the molecule. And when the standard method fails, which it will, have a fallback ready. I stop second-guessing my routes once I account for protecting group orthogonality early enough that it does not become a crisis mid-synthesis.