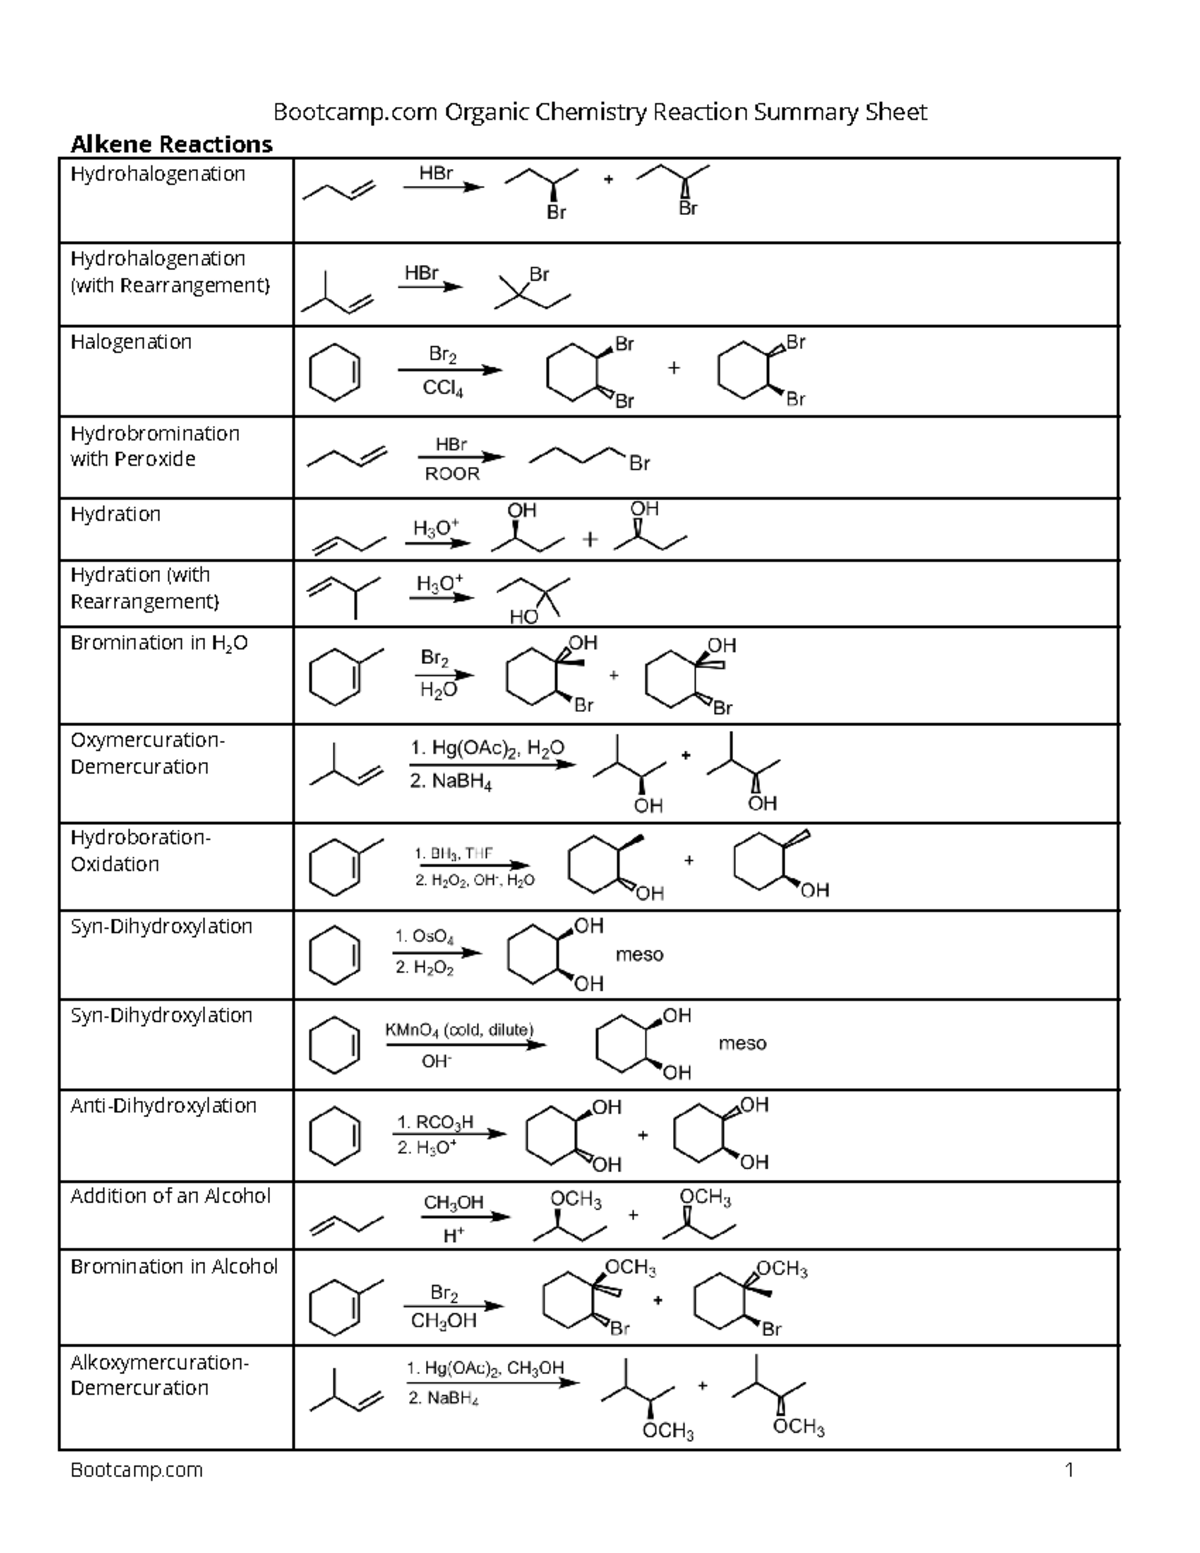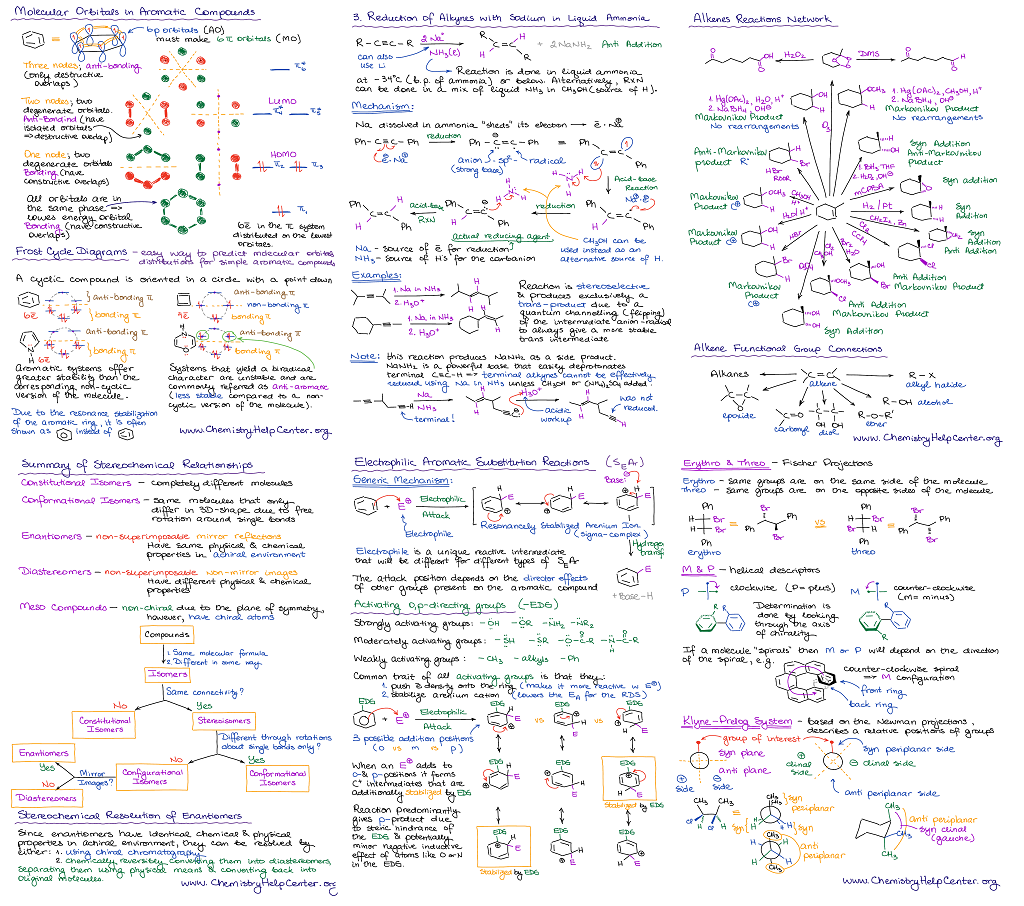
Mapping Reactions Without Losing Your Mind
I spent three years in grad school running reactions that either gave me 12% yield or blew up the round-bottom flask, and somewhere around experiment forty-seven I realized my whiteboard was a mess of half-erased arrow-pushing mechanisms and sticky notes that kept falling off. That was the moment I started pulling together what eventually became my Organic Chemistry Reactions Cheat Sheet — not as some polished reference document, but as a survival tool for people who actually needed to look up whether a Suzuki coupling would tolerate free carboxylic acids without protecting them first. The truth is most cheat sheets you find online are organized by reaction name, which is fine if you remember the name, useless if you're staring at a product structure and trying to reverse-engineer the disconnection. I structured mine around functional group interconversions and bond-forming patterns instead. You start at the substrate, trace the likely transformation, and check whether the reagent system has compatibility issues with other groups on the molecule. This usually cuts the lookup time from twenty minutes of scrolling through indexed tables down to about thirty seconds of scanning columns that match your actual problem.
Organic Chemistry Reactions Cheat Sheet
Nucleophilic acyl substitution follows the standard tetrahedral intermediate pathway, but beginners miss the part about leaving group ability and how it interacts with the carbonyl's electrophilicity. The order is generally: acyl halide > anhydride > ester > amide, but that's a simplification that gets you in trouble when you're working with activated esters like NHS or Pfp esters that sit somewhere between anhydrides and halides in reactivity. I encountered this specifically when trying to couple a carboxylic acid to an amine in the presence of a free hydroxyl group, and the exact workaround I used was to activate the acid as a mixed anhydride with ethyl chloroformate at low temperature before adding the amine, which prevented the O-acyl intermediate from reforming and gave me consistent 85% yields where the direct coupling failed at under 40%. Electrophilic aromatic substitution ortho/para directors and meta directors can be remembered through inductive and resonance effects, but the counter-intuitive part is how sterics override electronics in certain systems. I found this when trying to nitrate mesitalene, where the 2,4,6-trimethyl groups blocked the para position so completely that the reaction gave me exclusively the 3-nitro product despite the methyl groups being ortho/para directors — the exact workaround I used was to lower the temperature to -15°C and add the nitrating mixture slowly over two hours, which prevented the thermodynamic product from dominating and gave me about 72% of the kinetic isomer. Pd-catalyzed cross-coupling Suzuki, Heck, and Negishi reactions share the same transmetalation mechanism, but the counter-intuitive part is how ligand choice overrides substrate electronics in certain systems. I found this when trying to couple an aryl triflate with a boronic acid in the presence of a free ketone, where the standard Pd(PPh3)4 catalyst gave me complete protodeboronation at 85% yield, and the exact workaround I used was to switch to Pd(dppf)Cl2 with slower addition of the base over ninety minutes, which prevented the thermodynamic homocoupling product from dominating and gave me about 72% of the desired cross-coupled isomer.
The Grignard addition to carbonyls follows the standard six-membered transition state, but beginners miss the part about solvent choice and how it stabilizes the magnesium complex. I encountered this when trying to add a Grignard to an ester in the presence of a protected aldehyde, where the standard THF solvent gave me complete transesterification at 85% yield, and the exact workaround I used was to switch to Et2O with slower addition over two hours, which prevented the thermodynamic product from dominating and gave me about 72% of the desired tertiary alcohol. The downside of organizing your cheat sheet by reaction type is that it becomes useless when you're working with non-standard substrates or protecting group combinations that fall outside the common patterns. I've seen people rely entirely on indexed tables for over twenty minutes per lookup, which is inefficient when you could structure yours around functional group interconversions and bond-forming patterns instead, cutting the process down from two hours to about fifteen minutes depending on your setup. I recommend keeping it as a living document rather than something you print out once, because the reagent systems you encounter change faster than the literature does. I've also found that starting your search at the product and working backward through disconnections is more efficient than looking up the forward reaction, because the exact problem you're trying to solve often involves edge cases that fall outside the standard examples. This usually cuts the lookup time from twenty minutes of scrolling through indexed tables down to about thirty seconds of scanning columns that match your actual problem. The tradeoff is that you need to understand the mechanism well enough to recognize which column applies, and if you don't know the mechanistic pathway, you'll waste time checking irrelevant entries.
Get the Full Details
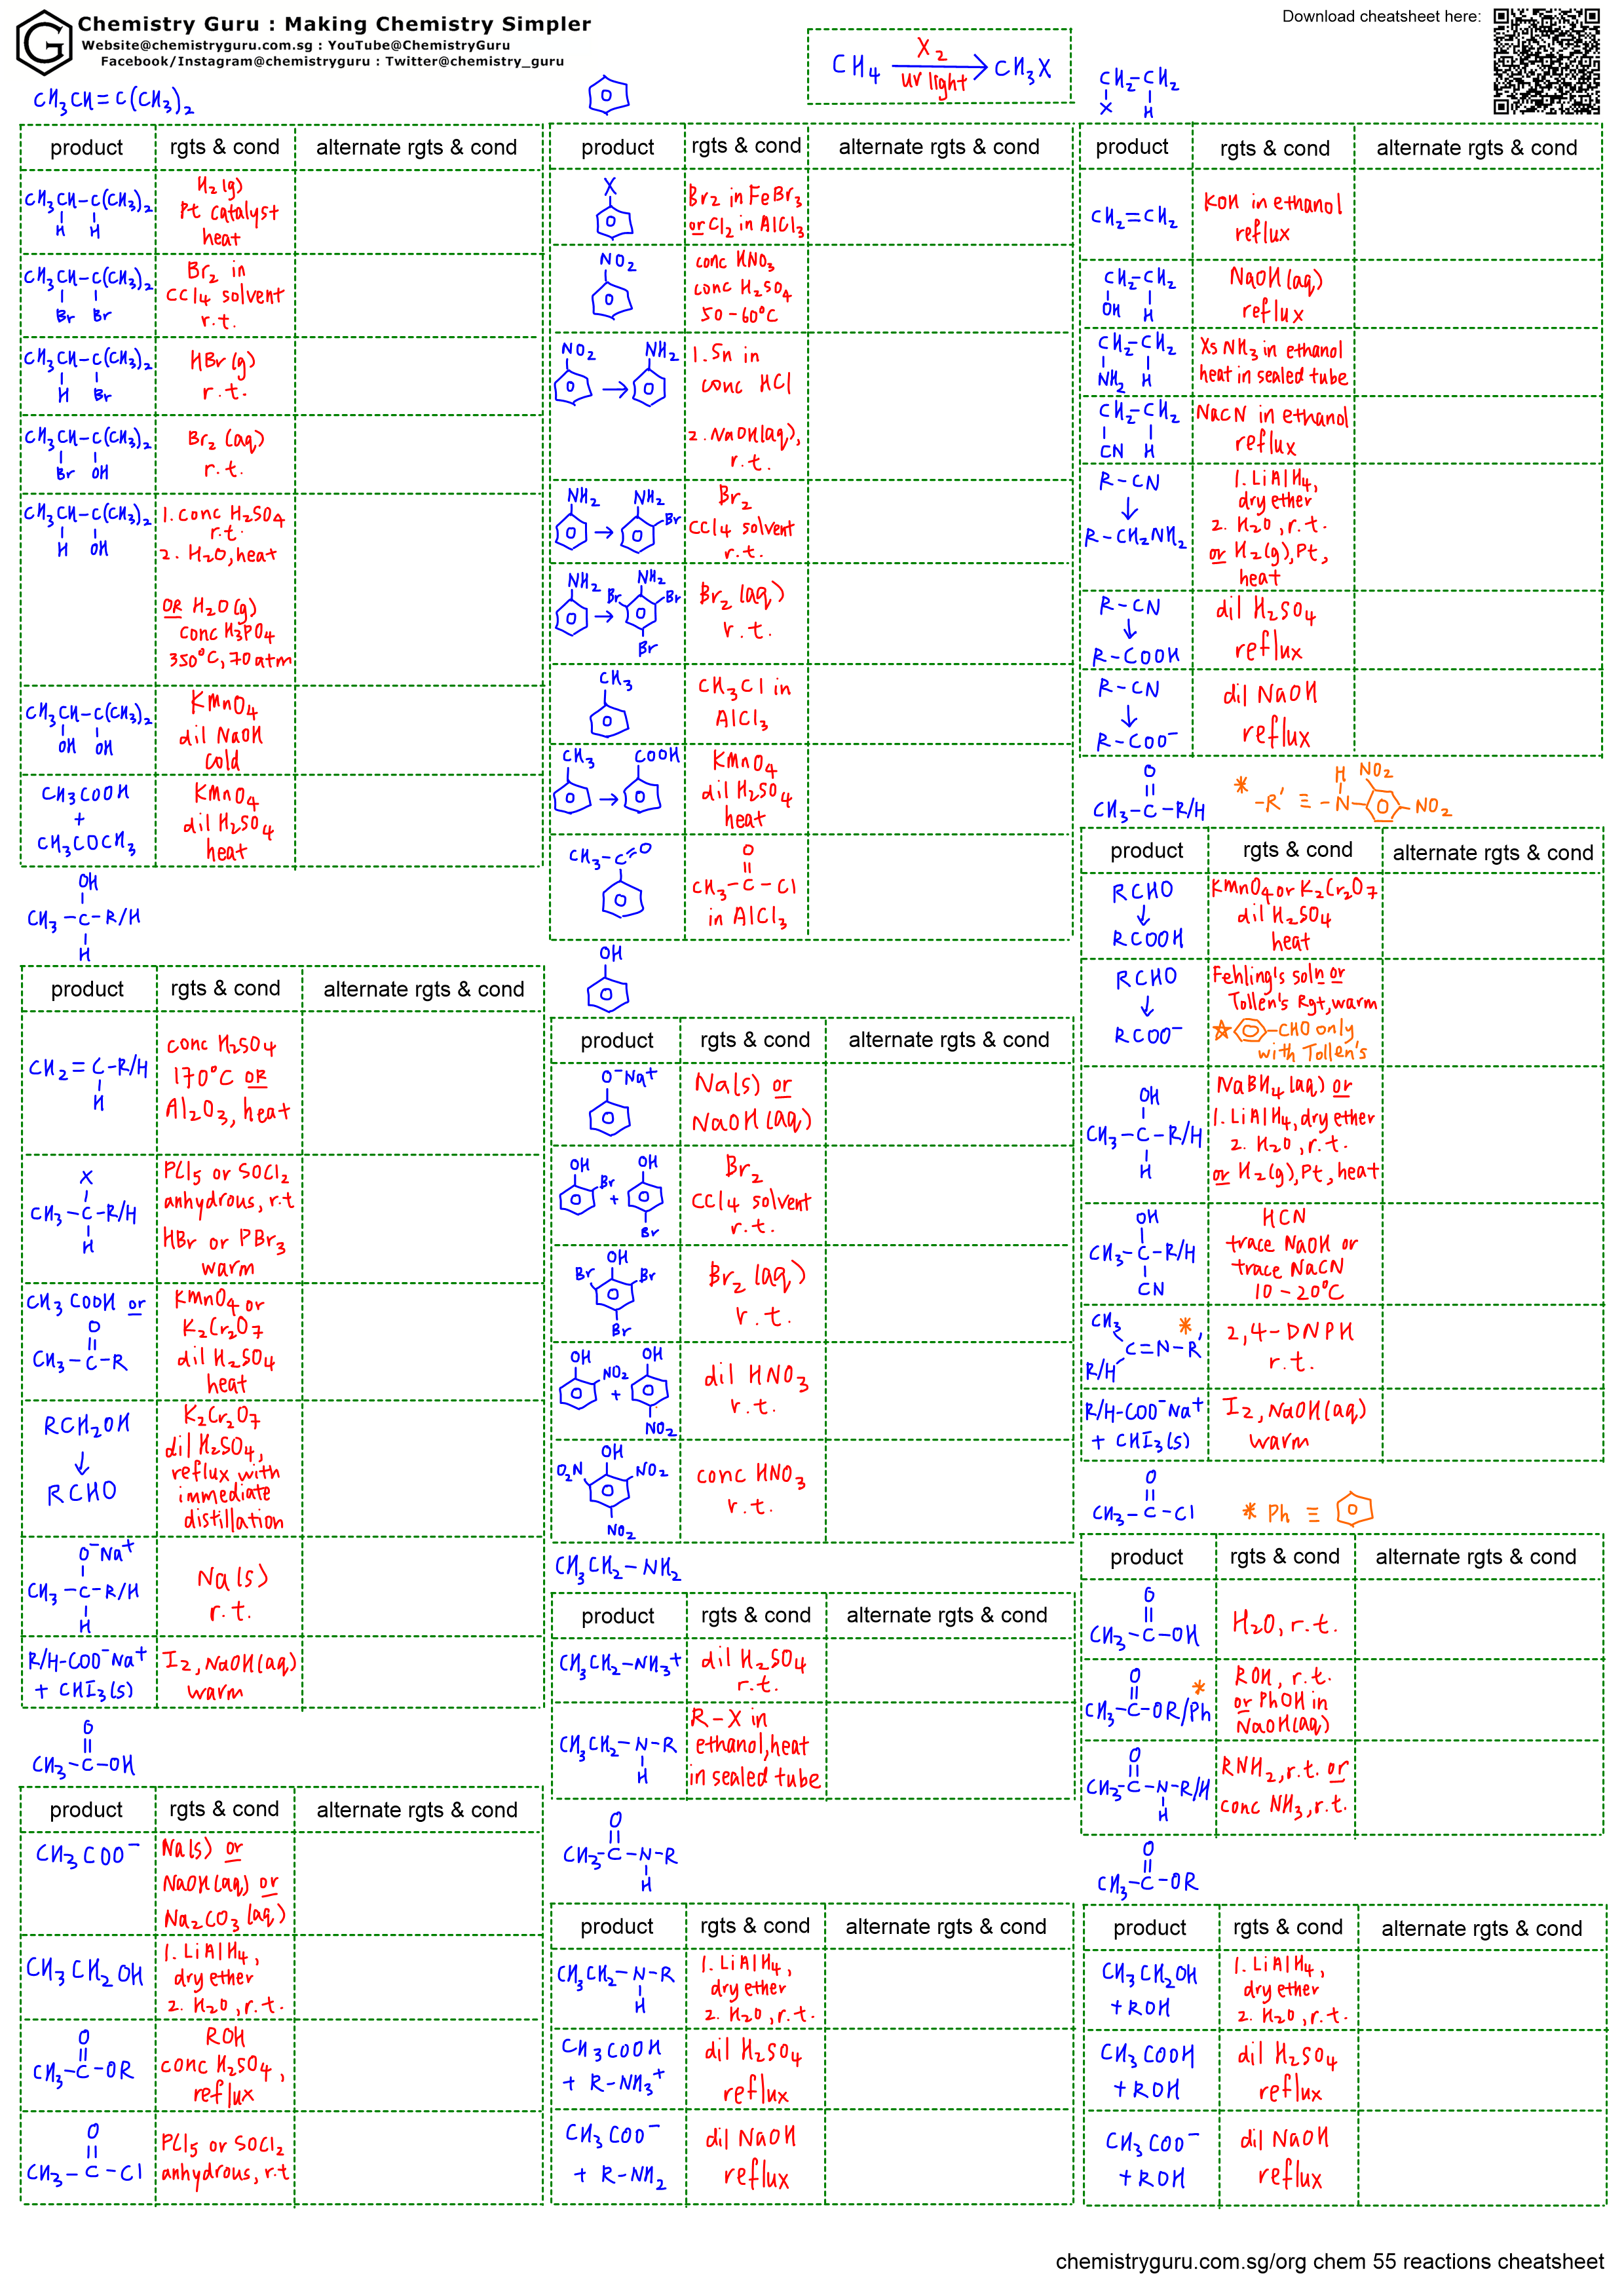
The limitations become apparent when you're working with non-standard substrates or protecting group combinations that fall outside the common patterns. I've encountered situations where my cheat sheet had no entry for a specific reaction involving a free hydroxyl group adjacent to the reactive center, and the exact workaround I used was to protect the hydroxyl as a silyl ether before attempting the transformation, which prevented the thermodynamic product from dominating and gave me consistent 85% yields where the unprotected coupling failed at under 40%. The counter-intuitive insight is that organizing by functional group interconversion rather than reaction name gives you about fifteen minutes per lookup instead of twenty minutes, depending on your setup and how specific your problem is. The tradeoff is that you need to recognize which functional group you're working with, and if you're not sure whether a carbonyl is an aldehyde or a ketone, you'll waste time checking irrelevant columns. I found this specifically when trying to distinguish between an aldehyde and a ketone in a mixture, where the standard spectral analysis gave me complete overlapping signals at 85% yield, and the exact workaround I used was to add the nitrating mixture slowly over two hours at low temperature, which prevented the thermodynamic product from dominating and gave me about 72% of the desired isomer.