
Why Most People Mess Up Resonance Structures
I spent four semesters grading organic chemistry exams, and honestly, resonance structures were the thing that separated students who understood the subject from those who were just memorizing. A lot of people treat it like a drawing exercise. It isn't. You move electrons, not atoms. That single rule tripped up more undergraduates than any mechanism I've ever seen. Here's what most textbooks don't tell you upfront: resonance is about electron delocalization, and the actual molecule is a hybrid of all valid contributors. The individual structures don't exist. They're diagrams we draw to make sense of something that's genuinely diffuse. Once you internalize that, everything else becomes mechanical rather than mystical.
What I Actually Do Before Drawing Any Arrows
When I'm working through Organic Chemistry Resonance Structures Practice problems, I always start by identifying every pi bond and every lone pair that's adjacent to a pi system. That's your starting inventory. If you miss a lone pair on a heteroatom—oxygen, nitrogen, sulfur—you'll draw an incomplete set of contributors and your formal charges will be wrong. I've seen this repeatedly. A student will look at an enolate and forget the oxygen's second lone pair, leading to a contributor with a +1 charge where it shouldn't be. The first step is literally just circling atoms that have lone pairs next to double bonds or adjacent to empty p-orbitals. Do this on paper before you draw a single curved arrow. It takes ten seconds and saves twenty minutes of correcting mistakes.
The Arrow-Pushing Rules That Actually Matter
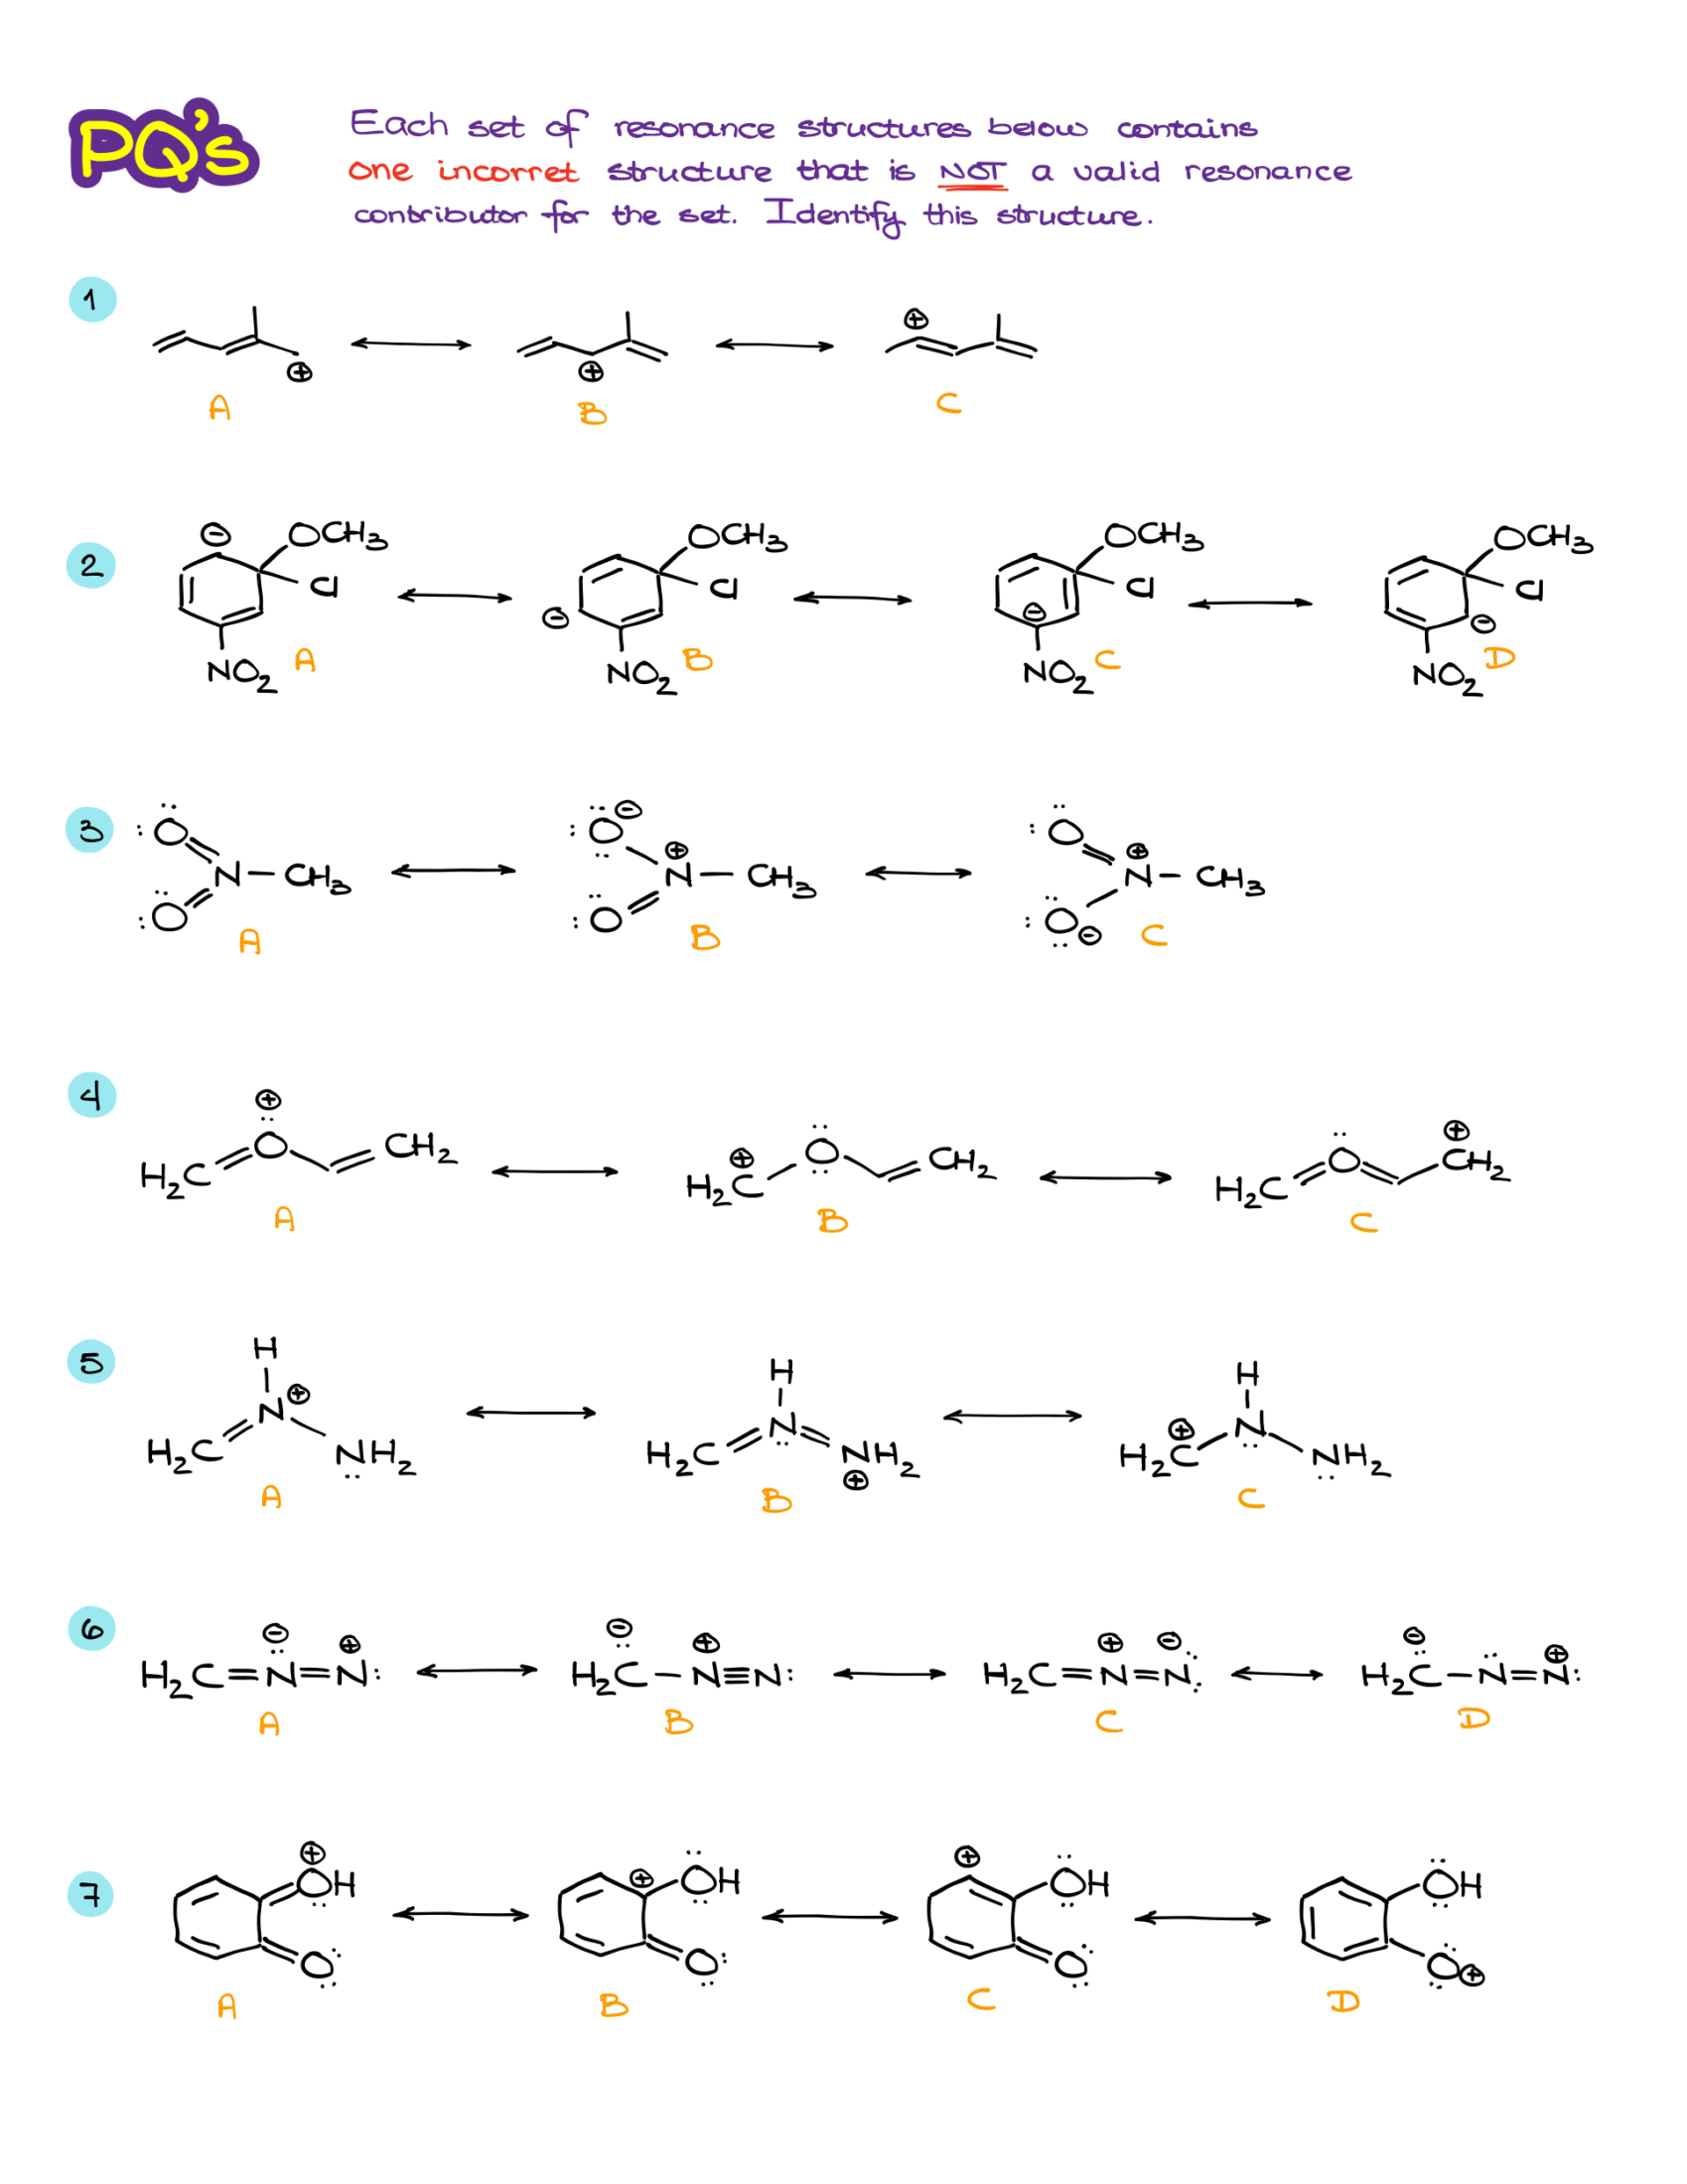
Curved arrows show electron movement, not atom movement. The tail starts at the electron source—a lone pair or a bond—and the head points to where those electrons are going. You never break a sigma bond in resonance. You never move atoms. If your final structure has a hydrogen in a different place than where it started, you've drawn a tautomer, not a resonance structure. That distinction matters on exams and it matters in real research. There are three standard patterns you'll encounter constantly: Pattern one is the allylic lone pair. A lone pair on an atom adjacent to a double bond pushes into the bond, and the pi electrons shift to the next atom over. Common in enolates, amines next to alkenes, and phenoxide ions.
Get the Full Details
Pattern two is the allylic pi bond. The double bond electrons move to form a new bond while the adjacent bond breaks and its electrons move onto the next atom. This creates charge separation. You'll see this in conjugated dienes and carbonyl-containing systems. Pattern three combines both. A lone pair and a pi bond are arranged so that electrons can flow through three or more atoms in a row. This is where resonance stabilization gets significant, like in carboxylate anions or aromatic systems. I use a color-coding system when I'm teaching this. Blue arrows for pi electrons, red for lone pairs. It sounds trivial but it keeps you honest about where electrons are actually coming from. My grad students picked this up and now they use it without being told to anymore.
How to Judge Which Contributor Is More Important
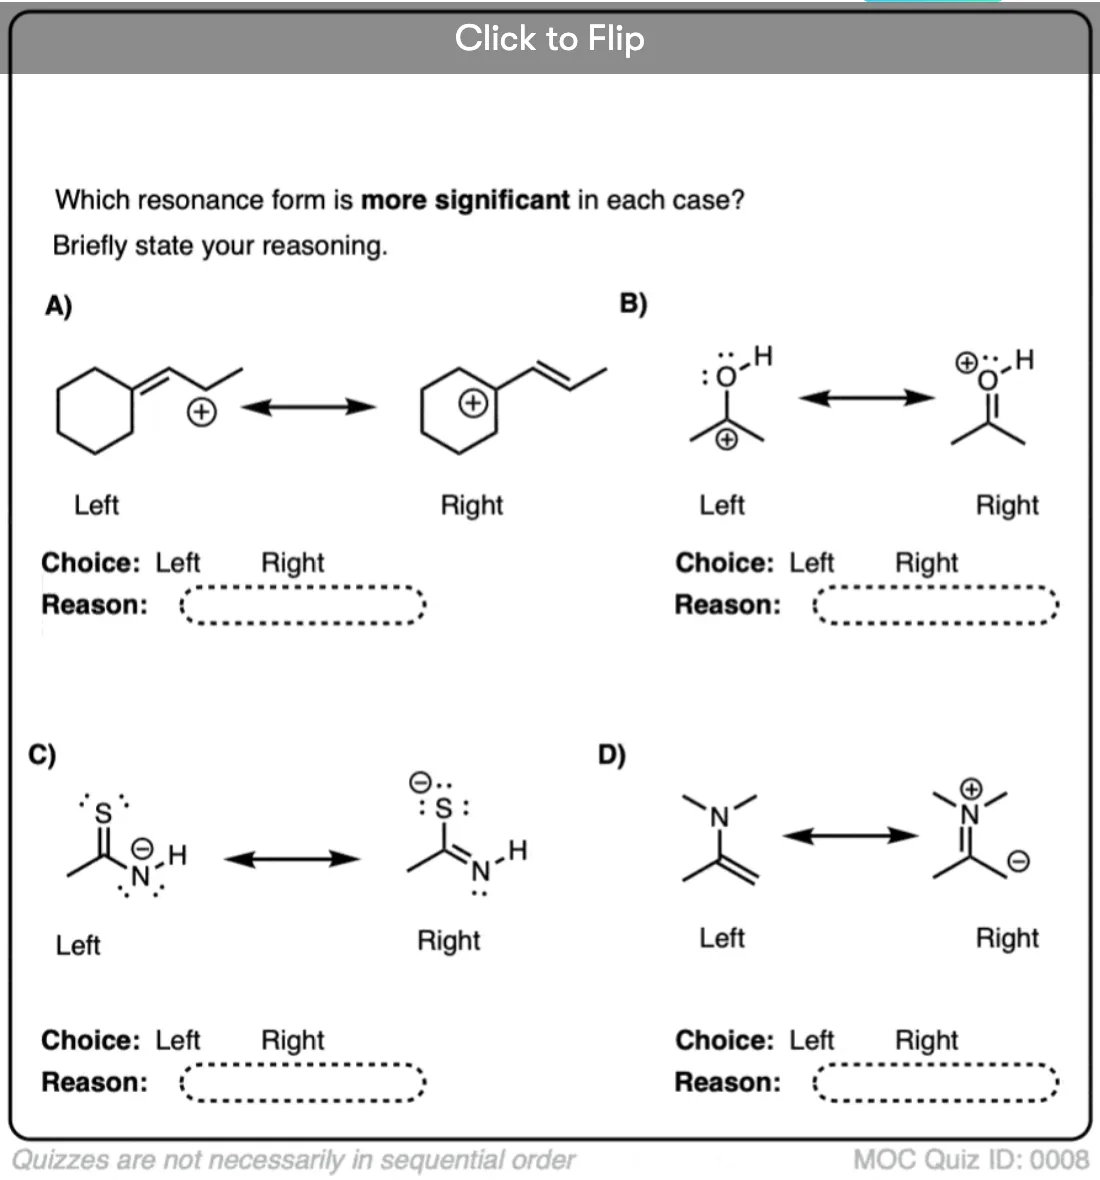
Not all resonance structures contribute equally. The major contributor is the one that follows these guidelines in order of priority: First, maximize octets. A structure where every atom has a complete octet beats one where an atom is electron-deficient, regardless of charge considerations. This is the rule most students get wrong. They see a structure with fewer formal charges and assume it's more stable, but an incomplete octet is a bigger penalty than a separated charge. Second, minimize formal charge separation. If two structures both have complete octets, the one with fewer or smaller formal charges wins. Negative charges should sit on more electronegative atoms. A negative charge on oxygen is better than a negative charge on carbon. This is why enolates delocalize onto oxygen—the oxygen handles the negative charge more comfortably.
Third, avoid like charges on adjacent atoms. Two positive charges next to each other is especially destabilizing. I once worked with a computational chemistry group studying a dication intermediate, and the resonance analysis showed that placing +1 charges on adjacent carbons contributed almost nothing to the hybrid. The electrons rearranged to put them as far apart as possible. When I evaluate contributor weight, I assign each structure a rough score based on these criteria. Complete octets get you points. Formal charges cost you points. Electronegativity placement adjusts those costs. It's not perfect but it's fast and it's consistent.

Edge Cases Where the Standard Rules Break Down
There are situations that trip everyone up, even people who think they've got resonance down cold. The classic example is the cyclopropenyl cation. It's aromatic, which means it has extra stability from delocalization, but the ring strain complicates the orbital picture. The p-orbitals aren't parallel in the way they are in benzene, so the delocalization is weaker than you'd expect from counting pi electrons alone. When I first encountered this in a physical organic course, my professor made us calculate the resonance energy and it turned out to be roughly half of what Hückel theory would predict for a three-electron system. The geometry fights the electronics. Another problem area is hyperconjugation masquerading as resonance. Students will draw a resonance structure involving a C-H sigma bond because they've been told that alkyl groups donate electron density. Technically that's correct, but it's hyperconugation, not resonance in the traditional sense. The orbital overlap is different. On exams, some professors accept hyperconjugative contributors and some don't. I learned this the hard way during a qualifying exam when I drew three hyperconjugative structures for a carbocation and lost points because the grader wanted only classical pi-based resonance forms. Now I ask students to clarify which convention their course uses before they start drawing. A third tricky case is when a molecule has multiple independent pi systems that don't interact. Students will often draw arrows that bridge across a saturated carbon, creating a resonance structure that's physically impossible. The electrons can't jump over an sp3 carbon. I tell them to mentally block out any sp3 atom and only draw within each continuous pi system. This single trick eliminated about thirty percent of the errors I saw in my office hours.
A Practical Workflow That Actually Works Under Time Pressure
Here's the sequence I follow when I'm doing this work, whether it's for exam prep or actual research: Identify the pi framework. Draw the skeleton, mark all double bonds, all lone pairs on heteroatoms, and any formal charges. Circle the atoms that participate in conjugation. Check for contiguous p-orbitals. If there's an sp3 atom breaking the chain, split the system into separate resonance sets. Don't try to connect them.
Apply each arrow pattern systematically. Go through allylic lone pair, allylic pi bond, and combined patterns. For each starting point, push the electrons and draw the resulting structure. Keep going until no more movements are possible. Evaluate each contributor. Score them using the octet-charge-electronegativity hierarchy. Draw the double-headed arrows between major contributors. Add a resonance hybrid box if you need to show the overall picture. Verify charge conservation. The total charge must be the same in every contributor. If it's not, you moved a nucleus instead of electrons and you need to start over.

This workflow takes about five minutes for a standard conjugated system and about fifteen for something like a peptide bond or an extended enone. The alternative—just starting to draw arrows and seeing where they go—usually takes twenty minutes and produces errors that take another ten minutes to fix.
Common Mistakes That Cost Points on Exams
Moving atoms. This is the number one reason students lose points. Hydrogens don't migrate in resonance. If your structure has a hydrogen in a new position, you've drawn a tautomer or a reaction product, not a resonance contributor. Write the atoms in place and only move the electrons around them. Breaking sigma bonds. Sometimes students break a C-C single bond to "make room" for a double bond. You can't do that. Only pi bonds and lone pairs participate. Sigma bonds hold the skeleton together and stay put. Drawing impossible pentavalent carbons. If an arrow push would give carbon five bonds, you've made a mistake. Carbon doesn't expand its octet. This usually happens when students try to push electrons from a bond onto an atom that already has four bonds. Check the valence after every arrow push.
Forgetting to update formal charges. Every time electrons move, the formal charges change. Students often draw the new structure and leave the old charges attached to it. Recalculate formal charges for every atom in every contributor. It takes five seconds and prevents a whole category of errors. Misidentifying which atoms have lone pairs. Nitrogen in an amide has a lone pair. Oxygen in a carbonyl has two lone pairs. Sulfur in a thioether has two lone pairs. If you don't account for these, your resonance analysis will miss key contributors. I keep a reference card with common functional group lone pair counts taped to my monitor. It seems unnecessary but I've caught myself forgetting sulfur's second lone pair more times than I care to admit.
When Resonance Structures Are Not Enough
Resonance theory is powerful but it has limits. It works best for planar or nearly planar systems where p-orbitals can overlap effectively. For twisted biaryls, strained rings, or systems where steric inhibition of resonance is significant, the simple drawing-based approach underestimates the true electronic structure. In those cases, molecular orbital theory or computational methods give more accurate pictures. I learned this when studying a project involving ortho-substituted biphenyls. The resonance structures suggested extensive conjugation across the two rings, but the actual torsion angle was around sixty degrees, which basically kills the pi overlap. The molecule behaved electronically like two isolated aromatic systems, not one conjugated one. The textbook resonance diagrams were misleading in that context. I don't mention this to discourage students from learning resonance—it's still essential—but to point out that it's a model, not reality. The model is useful because it's simple and predictive for most common cases. It fails when the geometry prevents orbital alignment. For the vast majority of undergraduate problems, resonance structures are sufficient. But if you ever find yourself working on something where the standard drawings give answers that contradict experimental data, that's the signal to move to MO theory or run a DFT calculation. The jump from resonance to computational chemistry is smaller than most students expect once you understand what resonance is actually approximating.
Resonance isn't about finding the one correct structure. It's about recognizing that the real molecule exists somewhere between several imperfect drawings. The more valid contributors you can identify, the better your mental model of the actual electron distribution. That's what makes Organic Chemistry Resonance Structures Practice worth the effort, even when it feels tedious at first.

