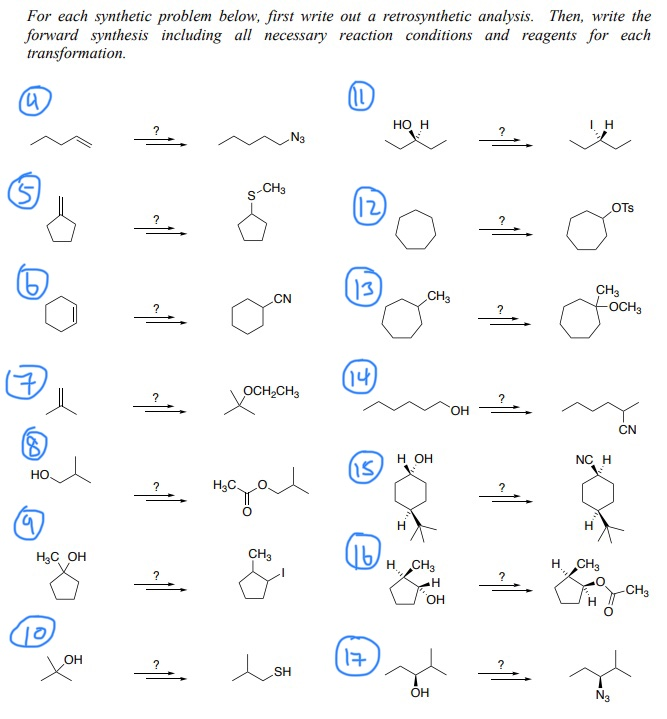
Working Through Retrosynthetic Analysis
Most people who ask for help with organic synthesis practice problems are stuck on the same things: they can identify functional groups but can't figure out how to disconnect a molecule into sensible precursors. Or they try to build forward from the starting material and end up with a mess of reagents that don't actually work together. I've seen this constantly. The core skill isn't memorizing reactions. It's learning to look at a target molecule and ask which bonds are worth breaking first. Disconnections should follow known mechanistic pathways, not random guesses. A good retrosynthetic step moves you closer to commercially available starting materials while keeping the strategy realistic.
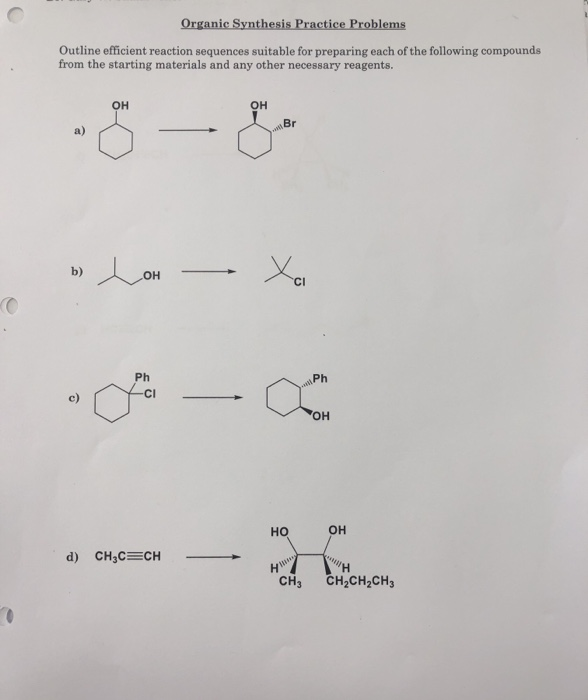
Organic Synthesis Practice Problems
Start by identifying the most strategically important bond in the target. Usually that's the bond connecting two fragments that would each come from simple precursors. Don't get distracted by the least substituted parts of the molecule. Focus on where you could form a carbon-carbon bond using something like a Grignard, aldol, Michael addition, or Wittig reaction. Here's a practical workflow I use when I'm working through a problem set: Write down the target structure clearly. Number the carbons so you can refer to them later. Look for disconnections that simplify the molecule by at least half. Check whether the resulting fragments are known, stable, and available. If a fragment would be unstable or impossible to isolate, your disconnection was wrong. Backtrack and try a different bond.
I remember working through a problem that asked for a synthesis of a bicyclic lactone with three stereocenters. The straightforward approach was to disconnect the ester linkage and assume a simple Robinson annulation. That route failed because the intermediate enone would undergo polymerization under the basic conditions needed. Instead, I disconnected the C-C bond adjacent to the ketone using a Michael addition with a stabilized malonate, followed by Dieckmann condensation. The malonate route gave clean yields and preserved stereochemistry because the Michael step set the critical stereocenter before cyclization. It took me two full attempts to see that the first disconnection was leading nowhere useful. One thing beginners consistently miss is that protecting groups are part of the problem, not an afterthought. If your synthesis requires a hydroxyl group to stay inert while you perform a reduction elsewhere, and you haven't planned a protecting group strategy early, you'll waste time troubleshooting failures later. Silyl ethers for alcohols and acetals for aldehydes are standard, but they add steps. Every protecting group costs you at least two operations: installation and removal. Factor that into your retrosynthetic count. Another counter-intuitive point: sometimes the best synthesis uses the least efficient reaction per step because it avoids a problematic intermediate. A 12-step linear synthesis with 90% yield per step gives you about 31% overall yield. A 15-step route with 75% per step gives you about 0.13% overall. But if the 15-step route avoids a purification nightmare or an unstable intermediate that decomposes during isolation, the 12-step route might give you less actual product in the flask despite the higher theoretical yield. I've lost whole batches to this. Once I stopped optimizing for step count alone and started accounting for practical isolation losses, my success rate improved noticeably.
Get the Full Details

When you're building a forward synthesis from a disconnection analysis, write out each reagent and condition with its known limitations. Don't assume a Grignard will tolerate an ester somewhere else in the molecule. It won't. Don't assume a Friedel-Crafts acylation will work on a ring already bearing a strong electron-donating group positioned ortho to the incoming electrophile. Sterics matter as much as electronics. I've wasted two days running a Suzuki coupling that failed because the boronic acid had decomposed from moisture exposure, and the protocol didn't mention storing it under argon.
Common Pitfalls and How to Avoid Them
Students often try to synthesize a target using a single clever reaction when two simpler, orthogonal steps would work better. Regioselectivity issues are the usual culprit. A Wittig reaction might give you the E-isomer when you need Z, or an aldol condensation might self-condense instead of cross-condensing. Planning for these outcomes upfront saves you from surprise. Atom economy matters in real lab work, even if textbook problems ignore it. A Corey-Chaykovsky epoxidation uses dimethylsulfonium methylide, which generates dimethyl sulfide as a byproduct. That smell lingers in the lab for days and requires proper ventilation. Safer alternatives exist, like using Oxone with a catalytic amount of a sulfide, but those require different workup procedures. Knowing the trade-offs between convenience and practicality is what separates a textbook answer from something you can actually run. Retrosynthetic analysis also breaks down when the target molecule has extreme steric bulk around the reactive center. I worked on a problem where the target had a quaternary carbon adjacent to the site where I needed to form a new C-C bond. Standard alkylations failed due to steric hindrance. Switching to a conjugate addition with a cuprate reagent allowed the bond formation to proceed, but only at low temperature with careful control of the addition rate. Rushing the addition caused homocoupling side products that were nearly impossible to separate by column chromatography.
For practice, start with molecules that have one clear disconnection point. As you get comfortable, move to targets with multiple functional groups and competing reactivities. Don't skip the stereochemistry. Drawing wedge and dash bonds correctly in your retrosynthetic steps forces you to think about whether a reaction is stereoselective or racemic. If a problem asks for a single enantiomer and your proposed route goes through a planar intermediate without any chiral influence, you haven't actually solved the problem. The best resource I've found for building this intuition is pairing every problem with its mechanistic explanation. Understanding why a reaction works is more useful than knowing that it works. When you hit a problem you can't crack, write out every possible disconnection and evaluate each one against known reactivity patterns. Eliminate the ones that violate basic organic chemistry principles. What remains is usually your answer.