
Working With Buffer pH: The Actual Process
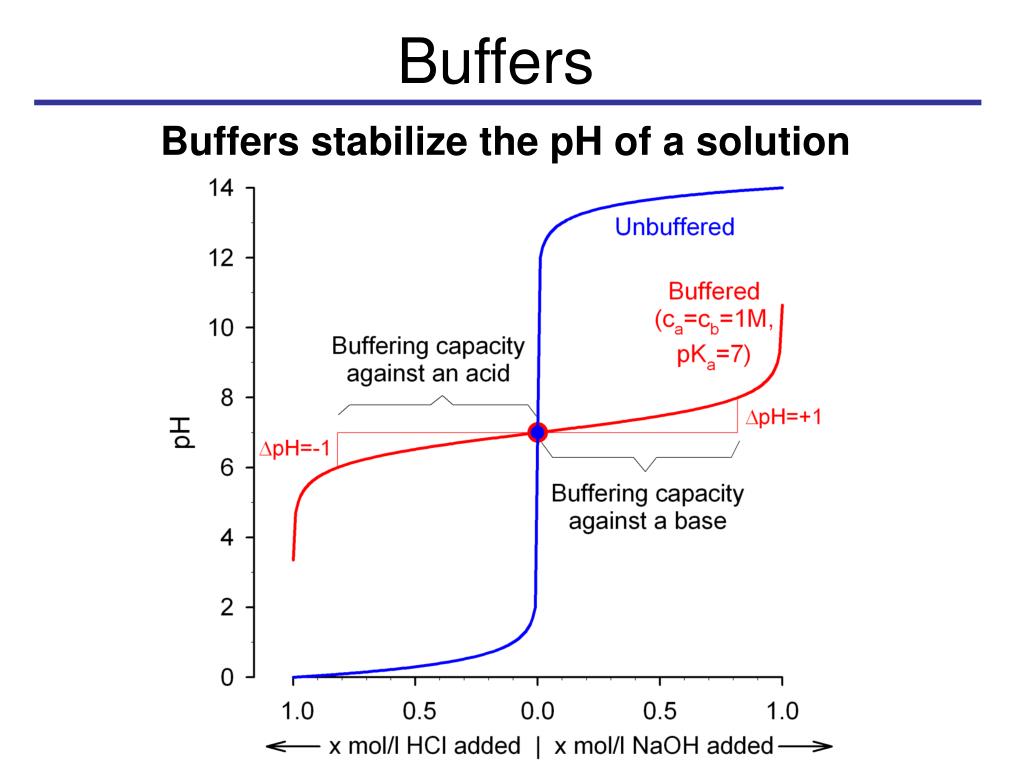
Buffer pH is one of those things that seems straightforward until you actually measure it and the number won't settle. You prepare a solution, stick a probe in it, and suddenly you're second-guessing everything you thought you knew about Henderson-Hasselbalch. Here's what actually happens and how to deal with it. The Henderson-Hasselbalch equation is your starting point, not your finish line. For a simple acetate buffer at pH 4.75, you'd mix acetic acid and sodium acetate in roughly equal molar amounts. The math gives you the theoretical value. Real measurements will differ. Temperature shifts, ionic strength, and the actual quality of your reagents all pull the number around. I've seen people calibrate their pH meter with fresh standards at 25 degrees Celsius, then measure the same buffer at 37 degrees and wonder why the reading jumped by 0.15 pH units. It's not a bad meter. The pKa of your buffer system shifted with temperature. phosphate buffers are especially sensitive to this. A 10-degree change can move your expected pH by nearly 0.1 units. If you're working with enzyme assays at body temperature, you need to account for that or your reaction conditions will be wrong from minute one.
The Practical Steps
Start with deionized water that's actually deionized. I can't stress this enough. Tap water will ruin your buffer before you even dissolve the first component. CO2 absorption from the air is another silent problem. Leave an open beaker of your buffer sitting on the bench for thirty minutes and the pH will drift, especially if you're working near neutral range. Carbonic acid forms. Your numbers shift. Cover it or work faster. When you dissolve your components, use a volumetric flask for the final volume, not a graduated cylinder. The difference between 100 mL and 100.5 mL in your stock buffer changes the concentration enough to matter when you're doing something like running a Western blot and consistency between blots is the whole point. I learned that the hard way after three independent experiments gave slightly different band intensities and I spent a week tracing it back to volume measurement error. For the actual pH adjustment, add your acid or base in small increments. I'm talking drops. Use a burette if you have one. The last 0.1 pH units take disproportionately more titrant than the first 0.5. Once you overshoot, you can't undo it without remaking a significant portion of the buffer. It's cheaper to waste 0.5 M NaOH drop by drop than to dump half your batch and start over.
Common Pitfalls That Nobody Warns You About
Tris buffers are a particular annoyance. Their pKa changes by about 0.03 pH units per degree Celsius. If you calibrate and measure at room temperature but run your experiment at 4 degrees or 37 degrees, your actual buffer pH is somewhere else entirely. I once spent two days troubleshooting an electrophoresis issue only to realize the running buffer's pH had drifted because the lab was air-conditioned that morning and the buffer was prepared the afternoon before. The gel ran fine, just not according to spec. Another one: ionic strength. The Henderson-Hasselbalch equation assumes ideal behavior. Real solutions don't behave ideally. At high concentrations, activity coefficients matter and your calculated pH will be off. If you're making a 1 M phosphate buffer, don't trust the textbook calculation alone. Measure it. The difference between the calculated and actual pH can be 0.2 to 0.3 units at that concentration. Old buffers degrade. Tris absorbs CO2 from the air over time and its pH drops. Phosphate buffers can precipitate if the concentration gets too high or the temperature changes drastically. I've opened a bottle of 1 M phosphate buffer that sat on the shelf for six months and found crystals at the bottom. The supernatant's pH was fine, but the concentration wasn't what the label said anymore. Toss it and make fresh. It's not worth the risk of a failed experiment.
Get the Full Details
Verification and Quality Control
Calibrate your meter before every use. Two-point calibration is the minimum. Three points if your work demands it. Use fresh standards. Old buffer standards degrade and your calibration becomes meaningless. NIST-traceable standards are worth the extra cost if you're publishing or running regulated assays. The cheap stuff from the discount supplier is fine for teaching labs but if you need defensible data, get the real standards. After you prepare your buffer, measure the pH and record it. Not the target pH. The actual measured pH. Future you will thank you when someone asks where the buffer came from and what its real pH was. I keep a logbook for this. It sounds excessive until you're trying to reproduce an experiment from six months ago and realize you wrote down 7.4 but the buffer was actually at 7.2 the whole time. For most routine work, this approach gets you into the right ballpark within five minutes of preparation time. If you need higher precision, account for temperature, measure after equilibration, and verify with fresh calibration standards each time. There's no shortcut around actually measuring the damn thing.


