Why Phase Changes In Science Matter When You Are Actually Running an Experiment
Most people learn about phase transitions in high school chemistry and think they understand it because they can recite that water boils at 100 degrees Celsius and freezes at zero. That is a textbook simplification that falls apart the moment you are working with anything other than pure H2O at sea level pressure. Phase Changes In Science is actually one of those areas where the theory and the practice diverge pretty significantly, especially if you are doing materials work, polymer processing, or anything involving alloy solidification. Let me walk through what actually happens and what trips people up, because I have seen enough failed batches to know where the pain points are.
Getting Real About Phase Changes In Science
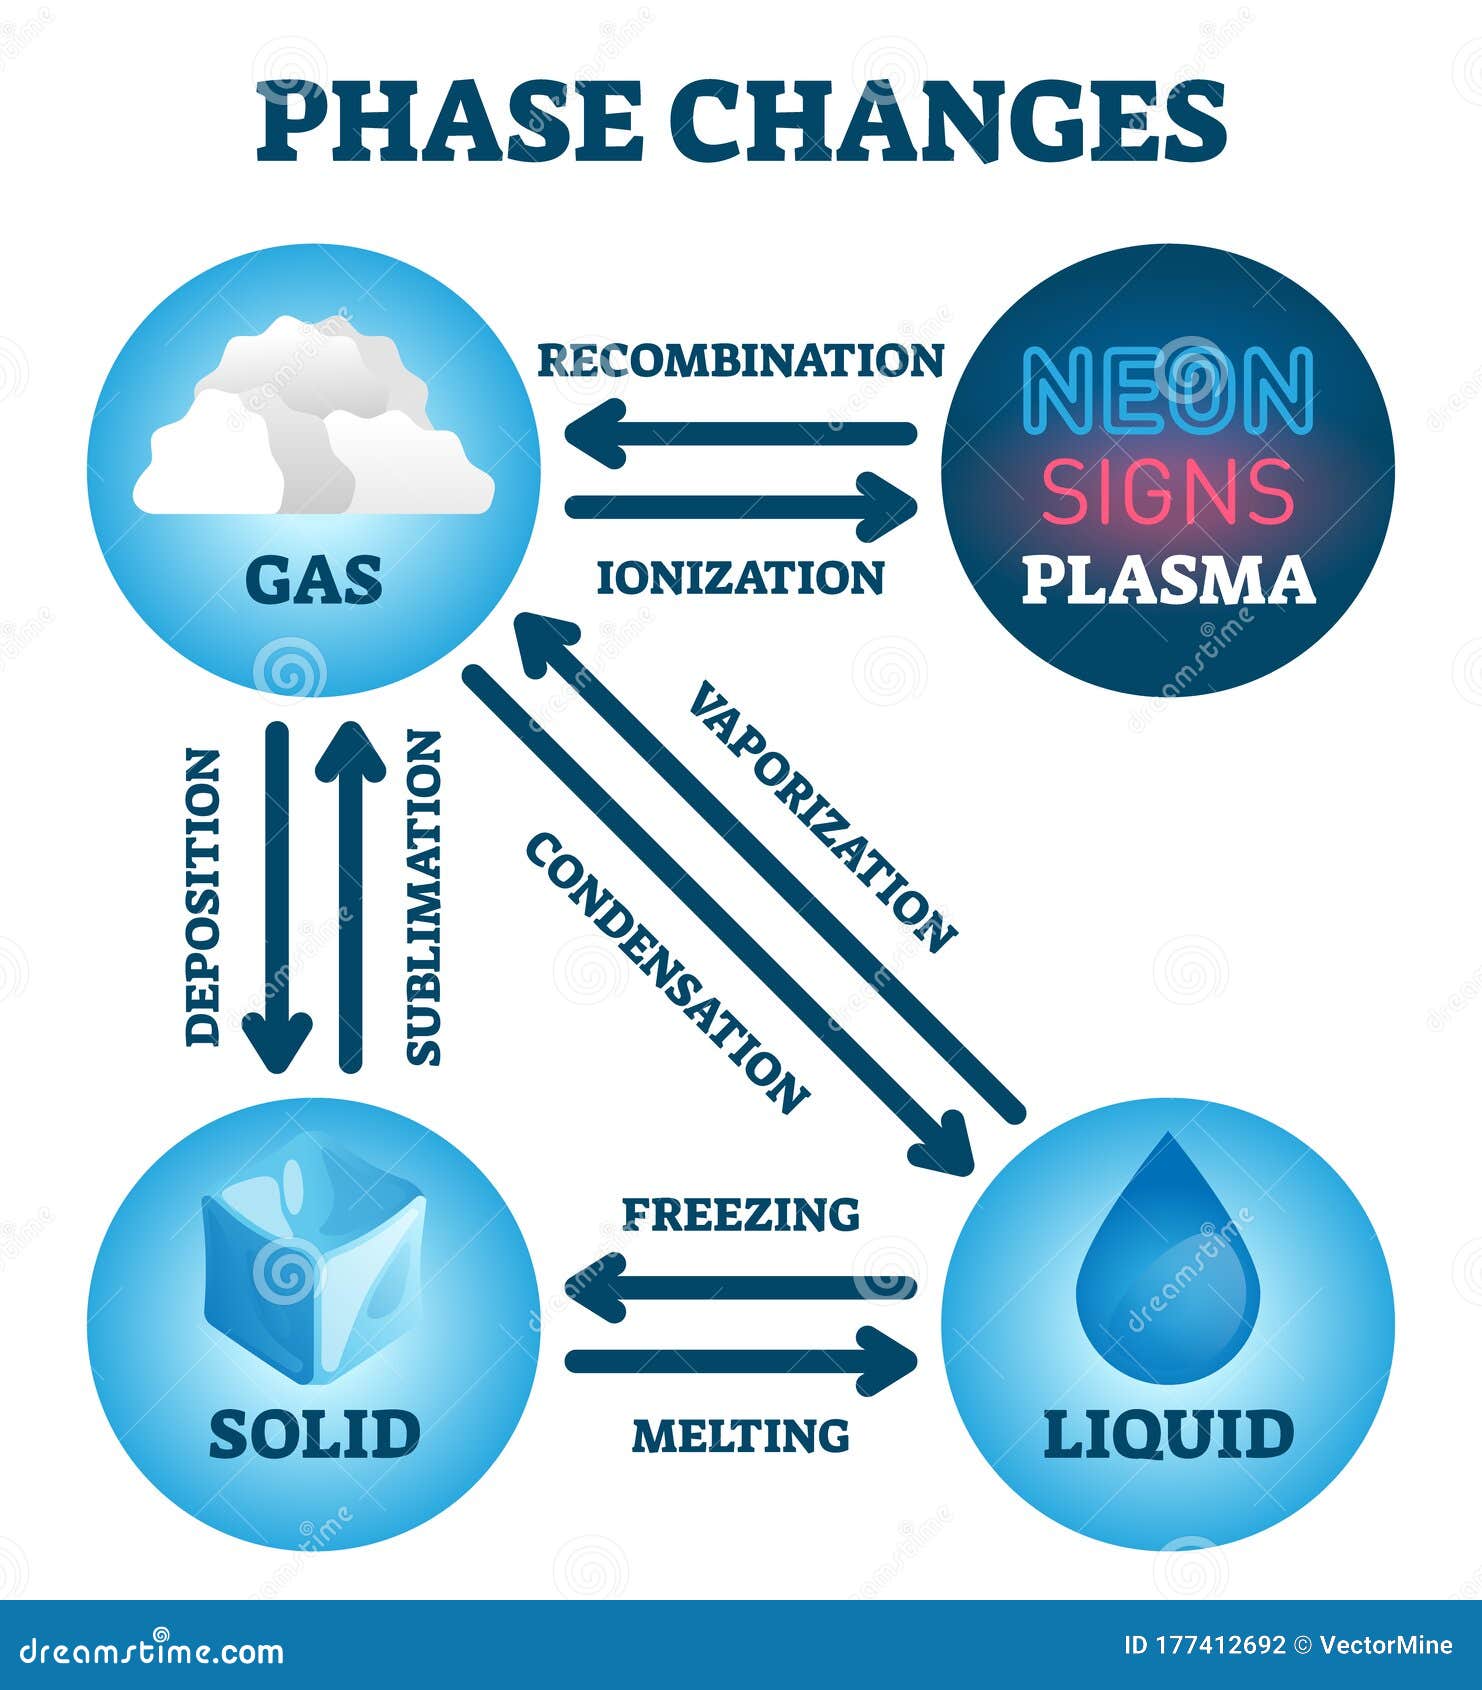
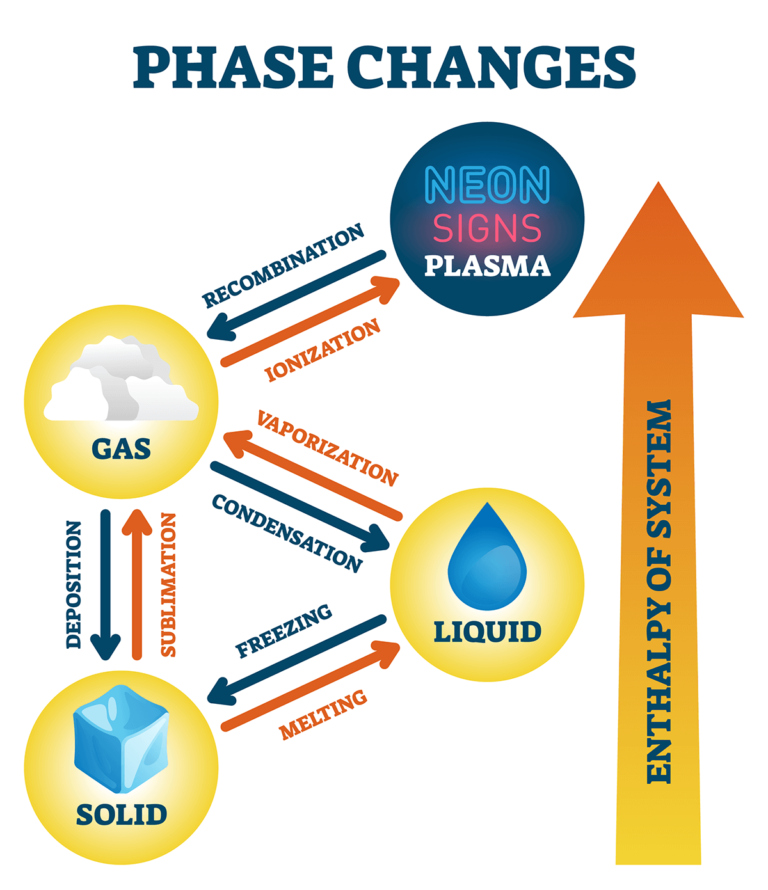
A phase change is fundamentally about a substance rearranging its molecular structure when energy crosses a threshold. The standard examples are solid to liquid, liquid to gas, and the reverse transitions. But the reality of working with this in practice involves supercooling, superheating, hysteresis, and metastable states that your textbook diagram completely ignores. When you cool molten glass fast enough, it does not crystallize into a solid — it forms an amorphous solid, essentially a liquid that became too viscous to flow on any meaningful timescale. That is a phase change that happened without ever passing through a true crystalline phase transition point. I learned that the hard way during a thermal annealing run that went sideways because I assumed the cooling curve would behave predictably. The practical implication is that temperature alone does not determine phase state. Cooling rate, nucleation sites, impurities, and pressure all shift where and when a transition actually occurs. If you are only monitoring temperature, you are missing half the picture. Here is a specific case that cost me three weeks of work and about four hundred dollars in wasted materials. I was working with a low-melting-point alloy for a casting project, and the spec sheet said the melting range was 71 to 78 degrees Celsius. I heated it to 85, poured it, and got a brittle casting full of voids. The problem was that the alloy had undergone partial recalescence — during solidification, the latent heat released by the early-forming solid phase briefly raised the local temperature, which then interrupted the rest of the solidification front. The voids were shrinkage porosity caused by uneven cooling, not trapped gas. The workaround was straightforward once I understood what was happening: I switched from a single pour into a room-temperature mold to preheating the mold to about 50 degrees Celsius and then letting the entire assembly cool in the oven with the heat turned off. That slowed the solidification enough that the recalescence event dissipated without disrupting the rest of the freeze front. The final casting was solid with no porosity.
The Practical Mechanics Most People Skip
Latent heat is the concept that causes the most trouble in real applications. It is the energy absorbed or released during a phase transition at constant temperature. When water boils, it sits at 100°C until every molecule has transitioned to gas, even though you are still applying heat. The energy is going into breaking intermolecular bonds, not raising temperature. This matters enormously when you are designing heating or cooling systems for phase change materials. If you are working with PCM — phase change materials — for thermal management, you need to account for the fact that during the transition, your material is effectively thermally inert. It will not get warmer or cooler until the phase change completes. This is why PCM is used in passive cooling applications: it absorbs a huge amount of energy while staying at a nearly constant temperature. The tradeoff is that once the material has fully melted or fully frozen, it loses that ability, and the temperature changes rapidly again. I have seen people design cooling systems around PCM without accounting for the finite capacity, and then wonder why the system overheated after six hours of runtime. The PCM was already melted and acting like a normal liquid at that point, which has a much lower heat capacity per unit volume than the latent heat it provided earlier. Another thing that beginners consistently get wrong is the difference between first-order and second-order phase transitions. First-order transitions, like melting or boiling, involve a discontinuity in entropy and volume — there is a measurable absorption or release of latent heat. Second-order transitions, like the ferromagnetic-to-paramagnetic transition at the Curie point, involve a continuous change in entropy but a discontinuity in heat capacity. There is no latent heat, no abrupt structural rearrangement, and the transition happens gradually over a small temperature range rather than at a sharp point. If you are characterizing a material and you see a smooth transition without a clear peak in your differential scanning calorimetry data, you may be looking at a second-order transition, not a measurement error.
Get the Full Details

Nucleation is another area where theory and practice diverge. Homogeneous nucleation — where a phase forms uniformly throughout a substance without any foreign surface — requires a much larger supercooling or superheating than heterogeneous nucleation, where the phase forms on impurities, container walls, or intentionally added seeding particles. In my experience, achieving reproducible results often comes down to controlling nucleation rather than controlling temperature. I once spent two days trying to get consistent crystallization in a polymer blend by adjusting heating rates alone. The breakthrough came when I added a tiny amount of a nucleating agent — just 0.1 percent by weight — which gave the crystals a consistent starting point and eliminated the batch-to-batch variation. The temperature profile stayed exactly the same. The difference was purely in the nucleation kinetics.
What Goes Wrong and How to Fix It
The biggest practical problem with phase transitions is thermal runaway during exothermic transitions. When a material releases latent heat during solidification, and that heat is not efficiently removed, the local temperature rises, which slows further solidification, which traps more heat, which creates a feedback loop. This is especially common in thick sections of casting or in large-scale PCM installations. The result is uneven microstructures, internal stresses, and sometimes complete failure of the part. The fix is usually straightforward: improve heat extraction during the transition phase. This might mean active cooling, increasing surface area through geometry changes, or using a material with higher thermal conductivity. It depends on your constraints, but ignoring the latent heat release during solidification is a reliable way to produce defective parts. Pressure effects are another frequent blind spot. The Clausius-Clapeyron equation describes how the equilibrium temperature between two phases shifts with pressure. For most substances, increasing pressure raises the melting point because the solid phase is denser than the liquid. Water is the famous exception — ice is less dense than liquid water, so increasing pressure actually lowers the melting point. This is why ice skates work. If you are working with water-based systems under pressure, assuming that higher pressure always means higher transition temperature will give you wrong answers. The direction of the shift depends on whether the substance expands or contracts during the transition, and you should check the density relationship before making assumptions. One more limitation worth stating bluntly: phase diagrams are equilibrium diagrams. They tell you what phases are stable at a given temperature and pressure when the system has had infinite time to reach equilibrium. Real processes are rarely at equilibrium. Rapid cooling can produce phases that do not appear on the standard diagram — martensite in steel is the classic example. It forms during rapid quenching and is a metastable phase that will not show up on an iron-carbon equilibrium diagram. If your process involves anything faster than slow furnace cooling, you are operating outside equilibrium conditions, and standard phase diagrams become predictive guides at best and outright wrong at worst. The workaround is to consult non-equilibrium diagrams or transformation kinetics data for your specific material, not just the equilibrium phase diagram.
There is also a practical limit to how well phase change materials perform in real-world thermal management. Over many cycle — melting and solidifying repeatedly — PCMs can degrade. Segregation can occur where different components of a eutectic mixture separate out, shifting the phase transition temperature and reducing latent heat capacity over time. I have seen PCM panels lose about 15 percent of their effective capacity after a few hundred cycles in a solar thermal storage application. The material was still functioning, but the performance drift meant the system design margins were eating away. If you are specifying PCM for a long-duration application, factor in degradation and oversize accordingly, or choose a material system known for cycle stability like hydrated salts with proper nucleating and thickening agents.

What to Actually Pay Attention To
If you are getting into this practically, the most useful skill is learning to read DSC — differential scanning calorimetry — data. A DSC curve shows you exactly when transitions happen, how much latent heat is involved, and whether your transitions are sharp or spread out over a temperature range. Sharp peaks indicate clean first-order transitions. Broad transitions suggest impurities, mixed phases, or kinetic effects. Learning to interpret those curves saves you from surprises later. Another practical tip: always measure your actual cooling and heating rates, not just the setpoint on your controller. PID controllers can oscillate, and thermal mass in your system means the actual sample temperature lags behind what the sensor reads. A thermocouple embedded in a block of material does not respond instantly. I use a separate reference thermocouple pressed against the sample itself whenever precision matters. The difference between the chamber temperature and the actual sample temperature during a phase transition can be several degrees, which is the difference between a clean transition and a messy one. The bottom line is that phase transitions are not as simple as crossing a temperature threshold. The rate at which you cross it, the presence of impurities or nucleation sites, the pressure conditions, and the thermal history of your material all matter. The textbook definitions are a starting point, not a working manual. Once you start accounting for the non-ideal behavior, things get more complex, but they also become predictable in a way that let you design around the problems instead of discovering them after the fact.