Getting Clean Plant Genomic DNA Without Losing Your Mind
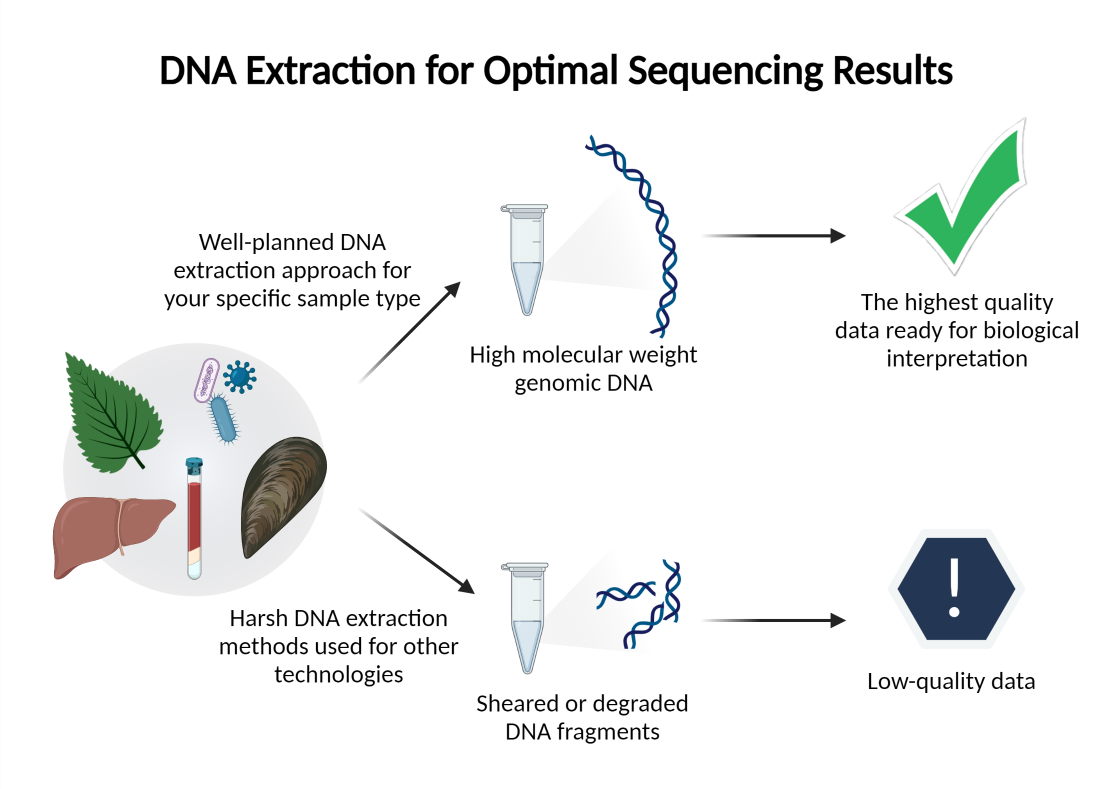
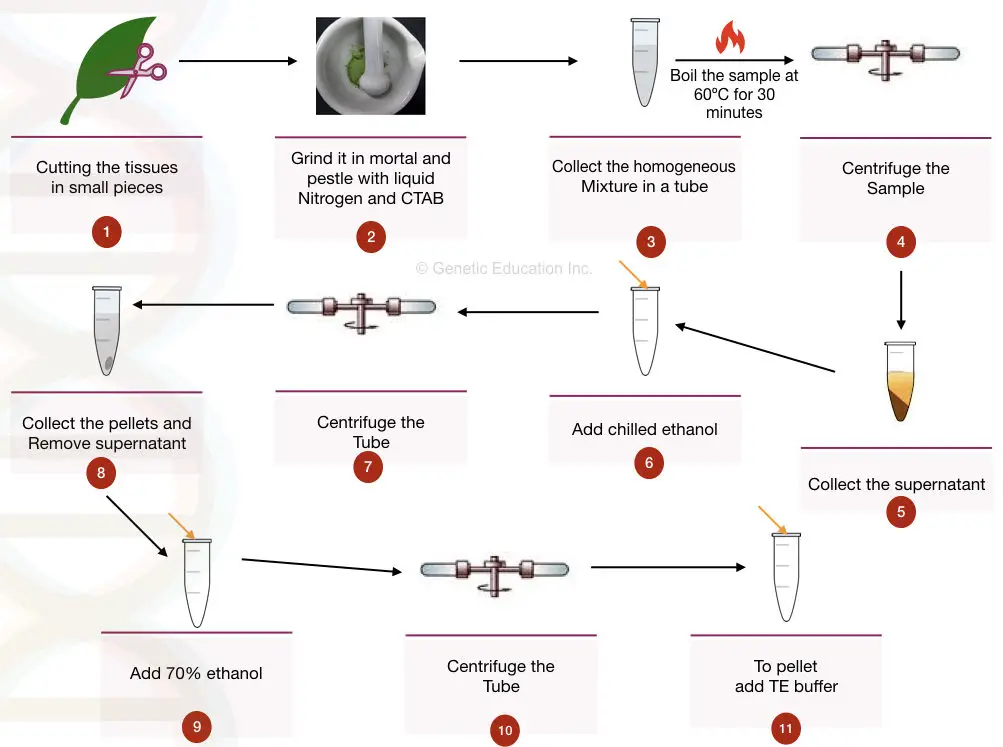
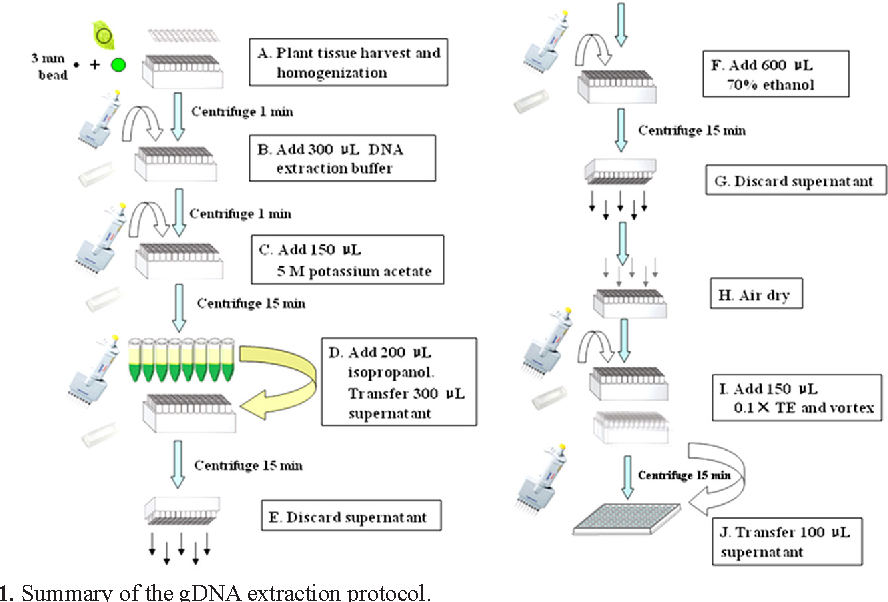
The standard CTAB-based extraction protocol from Integrated DNA Technologies is solid if you actually follow it, but most people cut corners and then blame the kit when their sequencing fails. I've done this probably three thousand times across various plant species, and the protocol itself isn't the hard part. What trips people up is the sample prep and the fact that plant tissue is inherently difficult because of polysaccharides, secondary metabolites, and phenolic compounds that co-precipitate with your DNA and wreck downstream applications. Here's how it actually works in practice. Start with young, healthy leaf tissue. Older leaves have higher polyphenol oxidase activity, and that enzyme turns your extract brown and useless within minutes of homogenization. Flash-freeze your samples in liquid nitrogen and grind them to a fine powder before they thaw. I learned this the hard way with a batch of Arabidopsis samples that I let sit on ice for too long while I sorted out my centrifuge settings. The resulting DNA was sheared to fragments under 200 base pairs, and my qPCR efficiencies were garbage because of it. Never let the tissue partially thaw before grinding. Add CTAB extraction buffer pre-warmed to 65 degrees Celsius directly to the ground powder. The buffer should contain 2% CTAB, 1.4 M sodium chloride, 20 mM EDTA, 100 mM Tris-HCl pH 8.0, and 2% PVP-40. The PVP binds phenolics. Skip it at your own risk. Incubate at 65 degrees for 30 to 60 minutes with gentle inversion every 10 minutes. Don't vortex. Vortexing shears high-molecular-weight DNA, and if you're doing anything requiring long contiguous sequences, you just made your sample unusable.
After the incubation, add an equal volume of chloroform-isooamyl alcohol (24:1) and mix by inverting 20 to 30 times. The phase separation matters here. Spin at 12,000 times g for 10 minutes at room temperature. You should see three distinct layers: white precipitate at the interface, the aqueous phase on top, and the organic phase below. Transfer only the aqueous phase to a new tube. Do not pipette any of the interphase material. Even a tiny amount of that white gunk means your DNA is contaminated with proteins and polysaccharides, and no amount of ethanol precipitation will clean it up properly. For the isopropanol precipitation step, add 0.6 to 0.7 volumes of isopropanol to the aqueous phase. Invert gently until you see the DNA spool. This usually takes 2 to 3 minutes. Spool the DNA onto a glass rod or pipette tip, then transfer it to 70% ethanol for a brief wash. Once it's washed, let it air dry for about 5 minutes. Don't over-dry it. If the pellet looks translucent and cracked, you've dried it too long and it will take forever to resuspend. That's not a minor inconvenience. I've wasted 45 minutes trying to resuspend a bone-dry DNA pellet that just wouldn't go into solution. Resuspend in TE buffer or nuclease-free water. If you're using a spectrophotometer, check your A260/A280 ratio. A clean prep should read between 1.8 and 2.0. If it's below 1.8, you have protein contamination. Run another chloroform extraction. If it's above 2.0, check your A260/A230 ratio. Values below 2.0 here mean residual phenol, chloroform, or salt carryover, which inhibits polymerases and restriction enzymes.
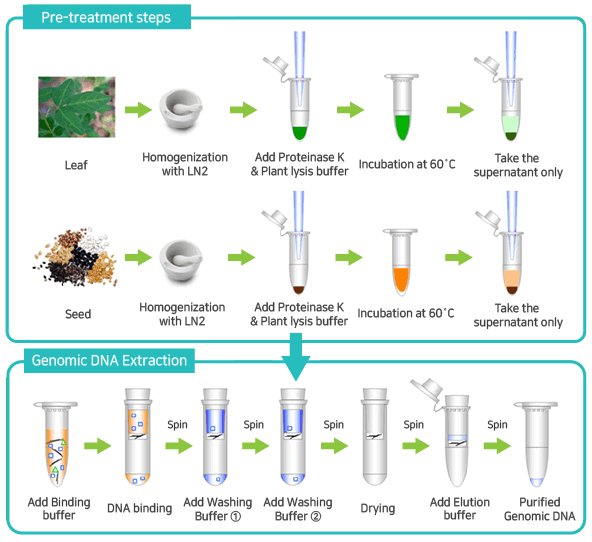
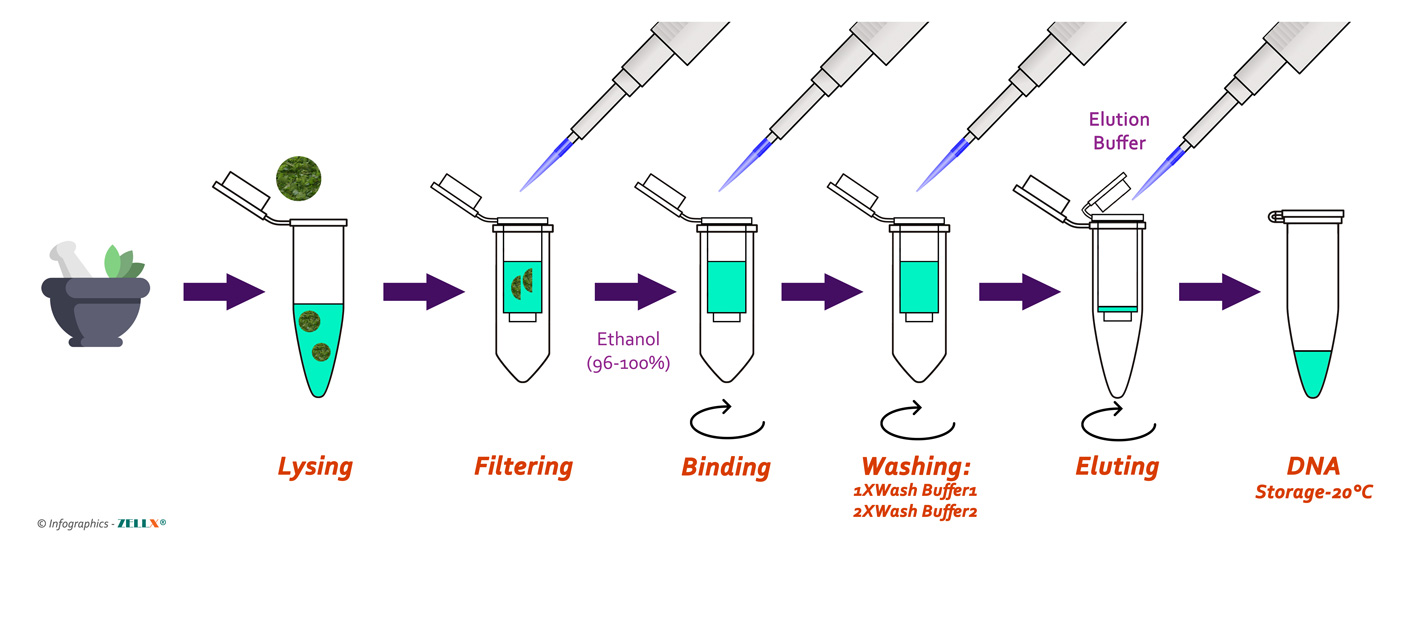
For species with high polysaccharide content like Cannabis or Eucalyptus, the standard CTAB protocol needs adjustment. I work with a lot of Cannabis samples, and the polysaccharide issue is real. These plants co-precipitate so much sugar that your DNA pellet looks gelatinous rather than fibrous. The workaround is adding an additional 0.1 to 0.2 volumes of 7.5 M ammonium acetate after the CTAB step and before the chloroform extraction. This helps precipitate polysaccharides selectively without losing DNA. It saves the sample every time. Another thing nobody tells you about: the overnight protocol. If your samples are difficult or you need maximum yield, you can incubate the CTAB mixture at 65 degrees Celsius overnight instead of 30 to 60 minutes. This is genuinely useful for tough tissues like woody stems or mature leaves with thick cell walls. The tradeoff is slightly more degraded DNA, but for standard PCR and genotyping applications, the difference is usually undetectable on an agarose gel. Integrated DNA Technologies sells a kits version of this called the NucleoBond Plant Plus Kit if you want a column-based approach, but honestly the CTAB method costs maybe 50 cents per sample versus $15 to $20 per sample with the kit. The column kits are cleaner for high-throughput workflows where you need consistent results across 96 samples, but for occasional extractions on a small number of samples, the CTAB method is faster and cheaper once you know what you're doing.
Get the Full Details

One more practical note about storage. Extracted DNA stored at minus 20 degrees Celsius will degrade over months due to nucleolytic activity from endogenous DNases that weren't fully inactivated during extraction. If you need long-term stability, store at minus 80 degrees or in solution with a final EDTA concentration of at least 1 mM. For routine PCR work, minus 20 is fine for a few months, but don't come back to the same aliquot more than three or four freeze-thaw cycles without expecting some degradation.
Common Failures and How to Fix Them
If your yield is low, the most likely culprits are incomplete cell lysis or loss during the isopropanol step. Hard-walled seeds and tissues require longer grinding times and possibly a second round of CTAB extraction. Some protocols recommend repeating the CTAB incubation on the same sample if the first pass gives poor yields. It works, but the yield from the second pass is usually half of the first. If your DNA won't dissolve, you almost certainly overdried the pellet. Add extra buffer and incubate at 55 degrees Celsius for 10 minutes with a gentle flick of the tube. Most of the time this gets it back into solution without needing to start over. If it still won't dissolve after that, the DNA may be irreversibly bound to polysaccharides, and you need to repeat the extraction with the ammonium acetate workaround mentioned above. The biggest mistake I see people make is assuming that all plant DNA extractions follow the same conditions. They don't. A protocol that works perfectly for Arabidopsis thaliana will give you terrible results for something like Ginkgo biloba or a cactus species. The tissue composition varies enormously, and you need to adjust salt concentrations, incubation times, and PVP amounts based on what you're working with. Read the literature for your specific species if you can find it. It saves a lot of wasted reagents and time.