Understanding Bond Polarity in Real Lab Work
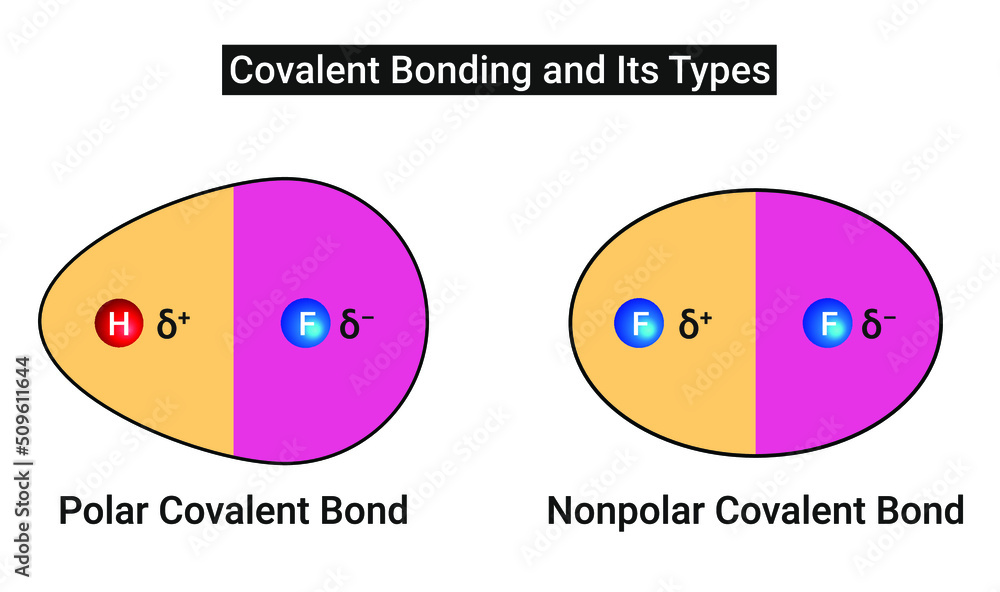
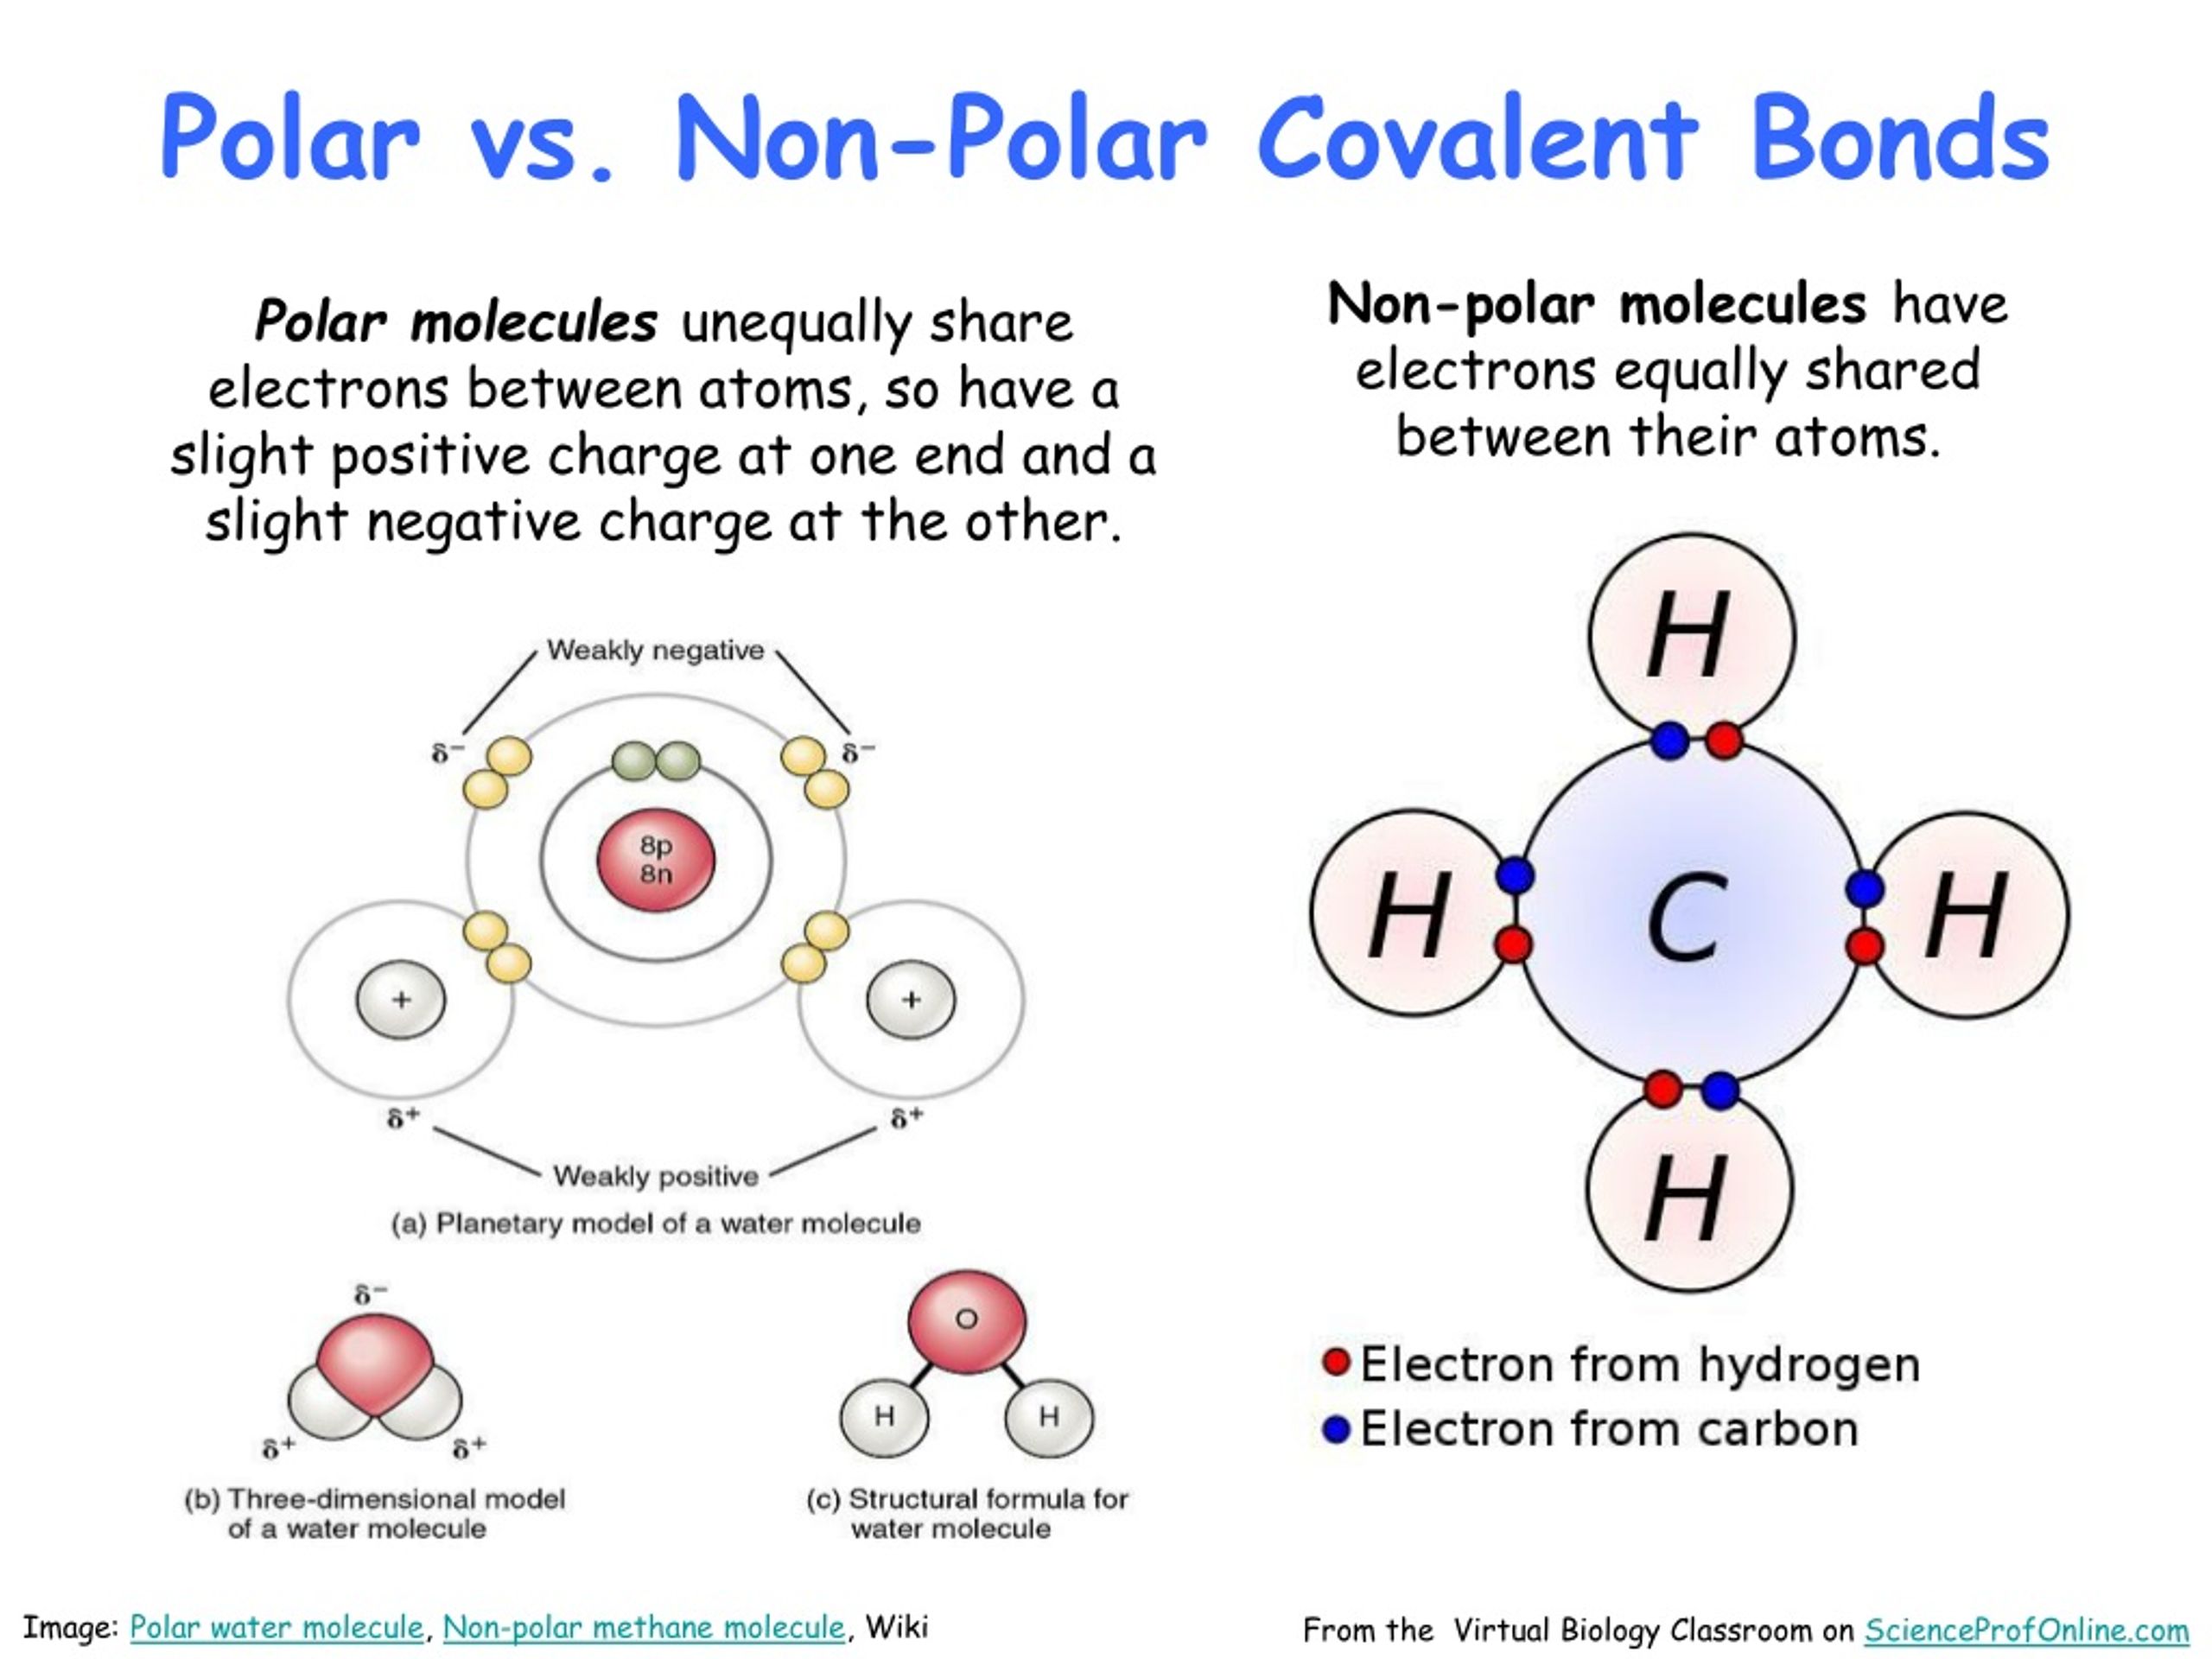
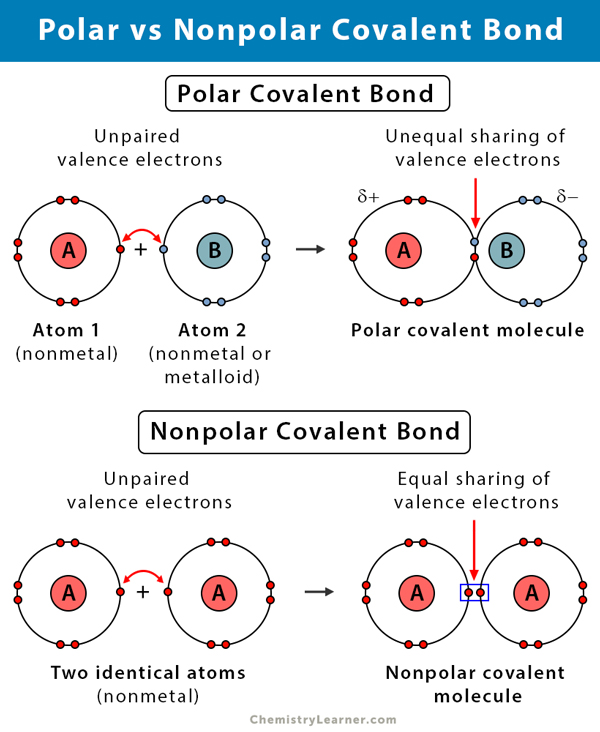
Most people learn about polar and nonpolar covalent bonds in a general chemistry class and think they've got it figured out. They remember "difference in electronegativity" and move on. That's usually enough for multiple choice tests, but it breaks down pretty quickly once you're actually dealing with molecules in solution or interpreting spectroscopy data. I ran into this head-on when I was troubleshooting solubility issues for a synthesis project a few years back. I had two compounds that looked identical on paper in terms of their functional groups, but one dissolved readily in water while the other refused to. The difference came down entirely to how the polar covalent bonds were distributed across the molecular geometry.The core concept is straightforward, even if the applications aren't. A covalent bond forms when two atoms share electrons. If the atoms have identical or nearly identical electronegativities, the electrons sit evenly between them. That's a nonpolar covalent bond. If one atom pulls harder on the shared electrons, the bond becomes polar, creating a dipole moment along that bond axis. I always tell people to stop thinking of polarity as a binary switch. It's a spectrum, and the actual behavior of a molecule depends on three things: the electronegativity difference between bonded atoms, the 3D geometry of the molecule, and whether those individual bond dipoles cancel out or reinforce each other. That third point is where most students trip up. Carbon dioxide has two polar C=O bonds, but the molecule is linear, so the dipoles cancel and CO2 is nonpolar overall. Water also has polar bonds, but it's bent, so the dipoles don't cancel and the molecule is polar. The bonds are the same type of thing, but the arrangement changes everything.
Polar Covalent Vs Nonpolar Covalent Bonds
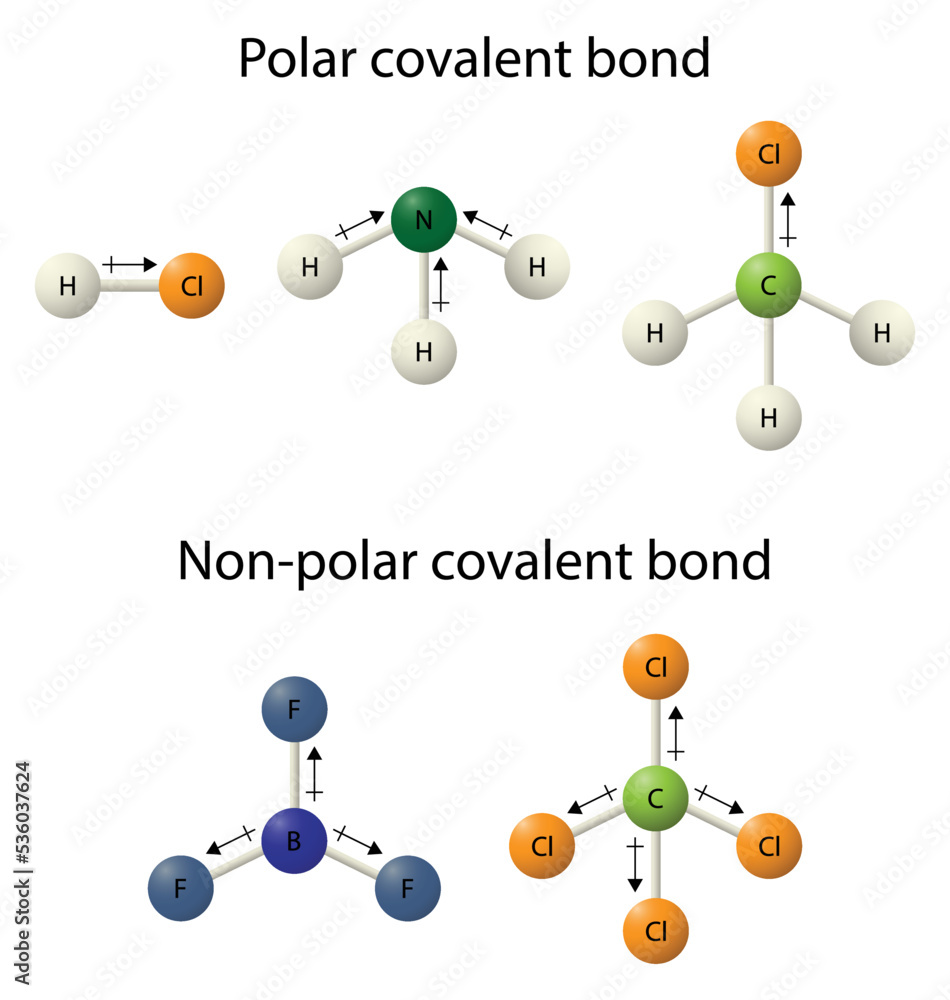
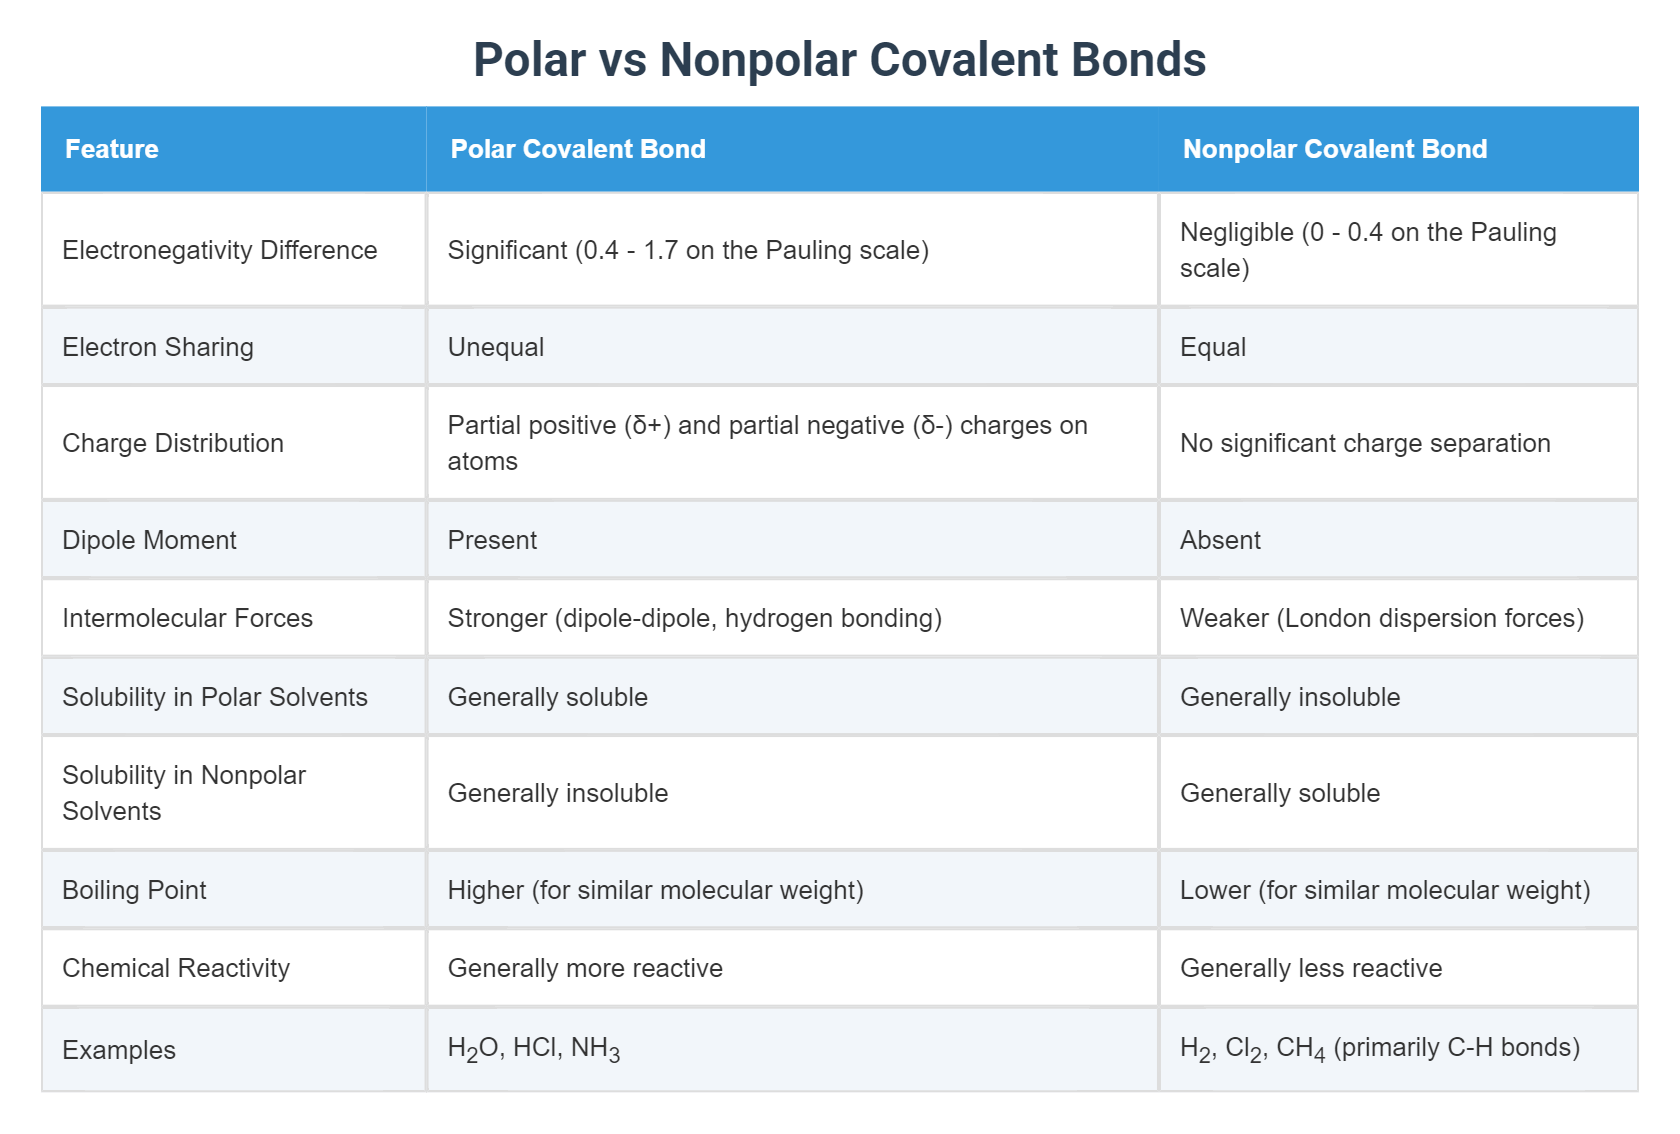
Here's how I actually determine which is which when I'm looking at an unknown compound or trying to predict its behavior. First, you need the electronegativity values. Pauling scale is the standard. You subtract the smaller value from the larger one for the bonded pair. If the difference is below 0.4, the bond is generally considered nonpolar covalent. Between 0.4 and 1.7, it's polar covalent. Above 1.7, you're usually looking at an ionic bond, though there are exceptions that depend on the specific elements involved and their oxidation states. But here's the part nobody emphasizes enough: the bond-level classification doesn't automatically tell you whether the whole molecule is polar. I spent way too long early in my career treating these as the same question. You have to do the vector addition of all bond dipoles. Draw the Lewis structure, figure out the geometry using VSEPR, then mentally add up those dipole vectors. If they sum to zero, the molecule is nonpolar despite having polar bonds. If they don't, it's polar. A practical example that stuck with me involved chloroform versus carbon tetrachloride. Both contain polar C-Cl bonds. CCl4 is tetrahedral and symmetric, so all four dipoles cancel and it's nonpolar. CHCl3 is also tetrahedral but the hydrogen breaks the symmetry, so there's a net dipole and the molecule is polar. This matters enormously if you're choosing a solvent for an extraction. CCl4 and chloroform behave completely differently in partition coefficients even though they look similar on paper.
When I'm working with real samples, I rarely rely solely on electronegativity tables. The numbers give you a starting point, but they don't account for hybridization effects, induction through sigma bonds, or the subtle influence of neighboring functional groups. In practice, I use a combination of calculated dipole moments from computational chemistry software and empirical data like dielectric constants or IR stretching frequencies. The C=O stretch in a polar environment shifts to a lower wavenumber compared to the same group in a nonpolar solvent. That's a direct read on bond polarity changes due to the surrounding field.
Get the Full Details
Common Pitfalls and Where the Simple Model Fails
The electronegativity difference method works well for straightforward cases, but it has real limitations. One issue is that it treats bonds in isolation. In conjugated systems or molecules with resonance, electron density gets distributed across multiple atoms in ways that a simple two-atom model can't capture. Benzene is a classic example. The C-C bonds all have the same electronegativity difference, yet the delocalized pi system creates regions of different electron density that affect reactivity patterns in ways basic polarity analysis misses. Another problem comes up with hypervalent molecules and transition metal complexes. The Pauling electronegativity values were derived primarily from main group elements, and applying them to d-block chemistry can give misleading results. I ran into this when modeling ligand binding affinities. The simple bond polarity calculations suggested one coordination geometry should be favored, but the actual experimental results pointed somewhere else entirely. The fix was to incorporate crystal field theory and look at the actual electron density distribution around the metal center rather than relying on bond dipoles alone. There's also the question of dynamic polarity. Bonds aren't rigid. Thermal vibration, solvent interactions, and conformational changes all shift electron distribution in real time. A molecule that appears nonpolar on paper might develop a significant transient dipole during a reaction event. This is especially relevant in enzyme active sites where the local environment can polarize bonds that would be nonpolar in solution.
For quick lab decisions, I usually fall back on solvent solubility rules and dipole moment measurements when available. If you need to know whether a molecule will interact strongly with water or partition into an organic phase, measuring its actual dipole moment with a dipole meter or calculating it with a program like Gaussian or ORCA is more reliable than any hand-calculation. The computational cost is low for small molecules, and the results tend to correlate well with observed behavior.
When Polarity Predictions Go Wrong
I should mention a case where my initial polarity assessment led me astray. I was evaluating a series of fluorinated surfactants for a formulation project. Based on electronegativity differences, I expected the perfluorinated chain to create a highly polar surface layer. The fluorine atoms are extremely electronegative, after all. The reality was the opposite. The C-F bonds are so tightly held and the chain so rigid that the surface ended up being low-energy and hydrophobic, almost Teflon-like. The individual bond polarity was high, but the collective molecular behavior was dominated by the low polarizability and tight electron packing of the fluorocarbon chain. This taught me to treat bond-level polarity as one input among many, not a prediction engine on its own. Molecular polarizability, surface area, hydrogen bonding capacity, and steric factors all compete with pure dipole effects. The most useful approach is to combine the electronegativity analysis with actual measured or computed properties rather than trying to derive everything from first principles. If you're studying this for an exam, focus on geometry and dipole cancellation. If you're using this in practice, measure what you can and model the rest. The theory gets you in the right neighborhood, but real chemistry rarely stays that tidy.