How Plasma Spectroscopy Actually Works In A Real Lab
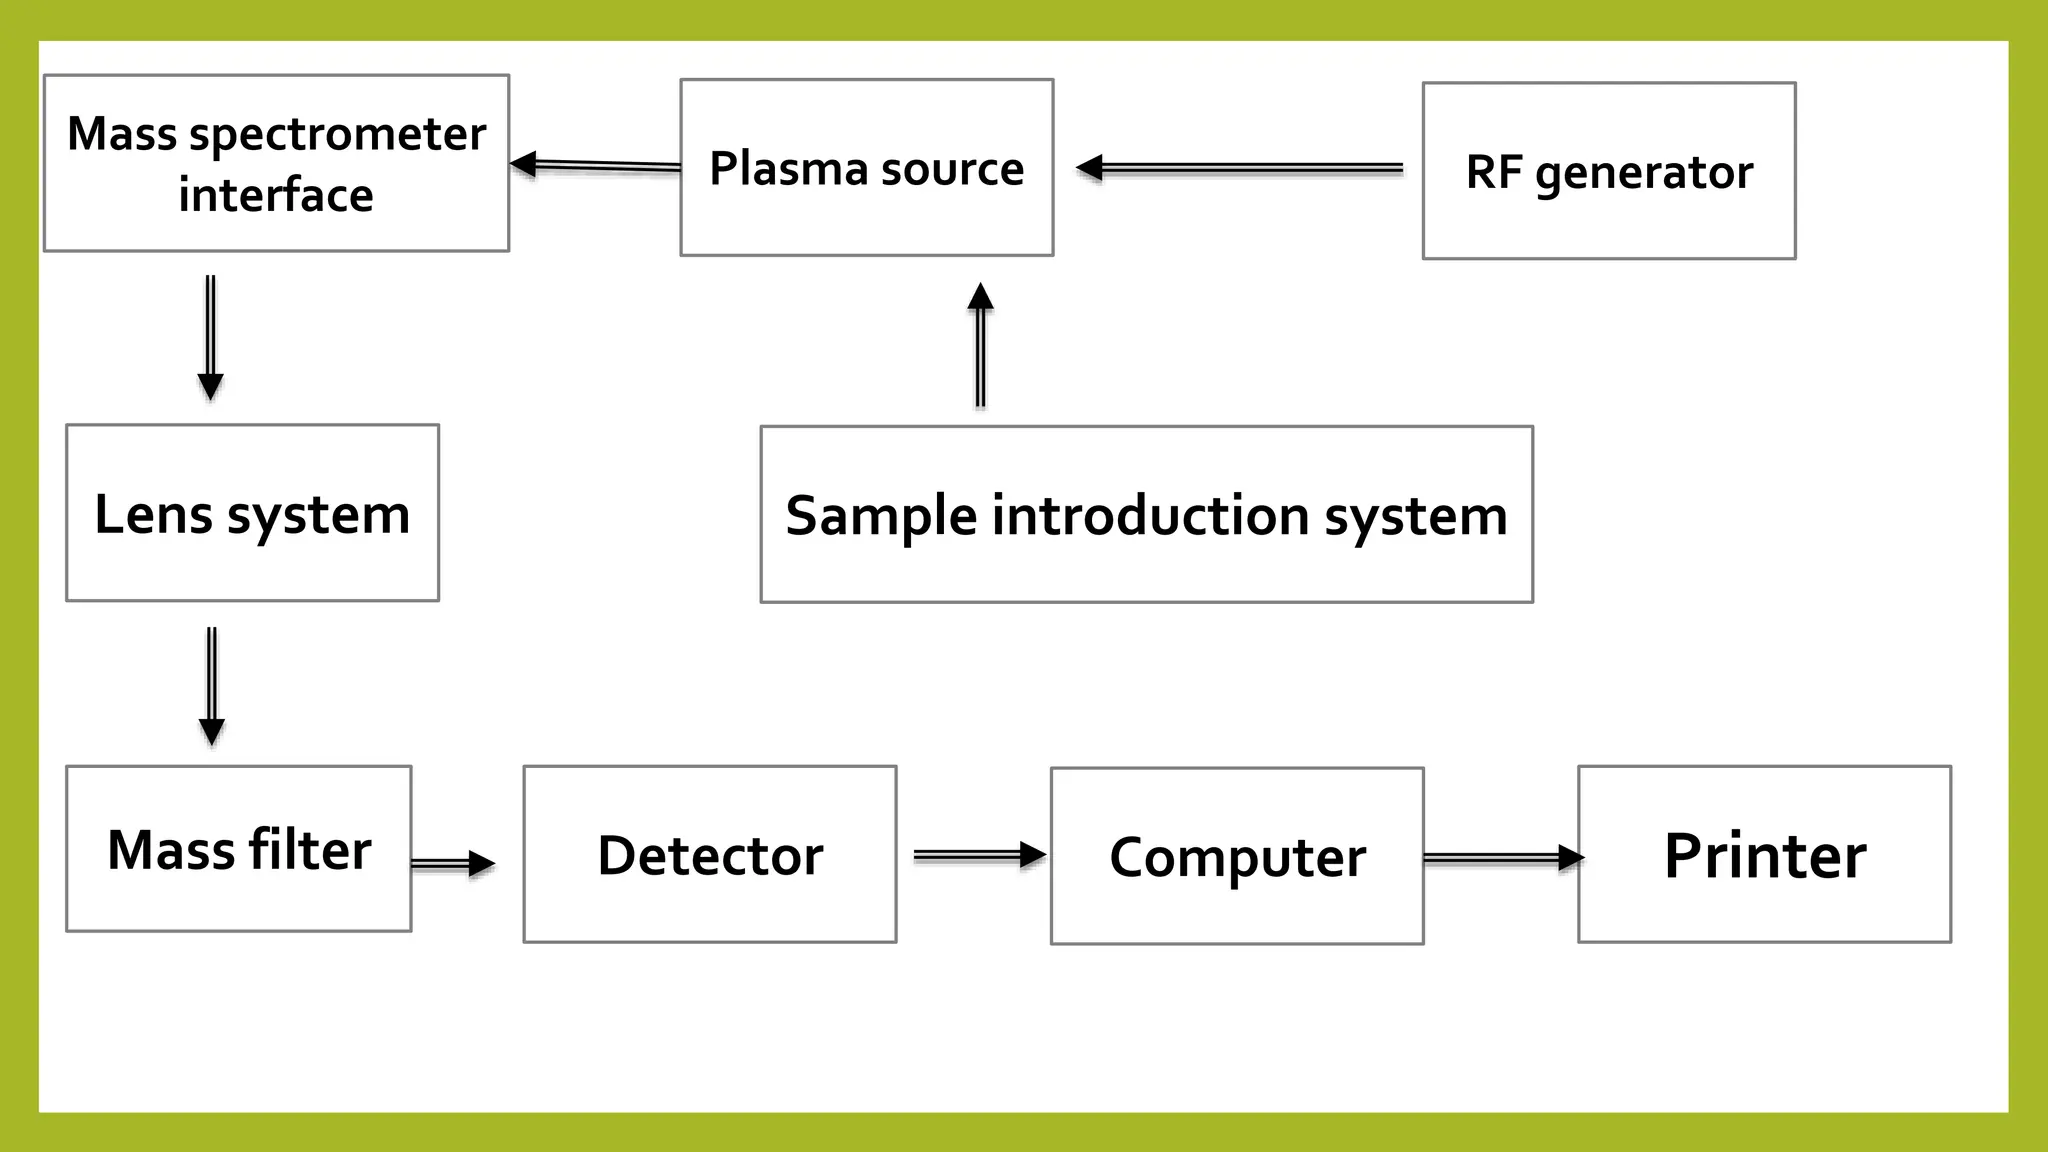
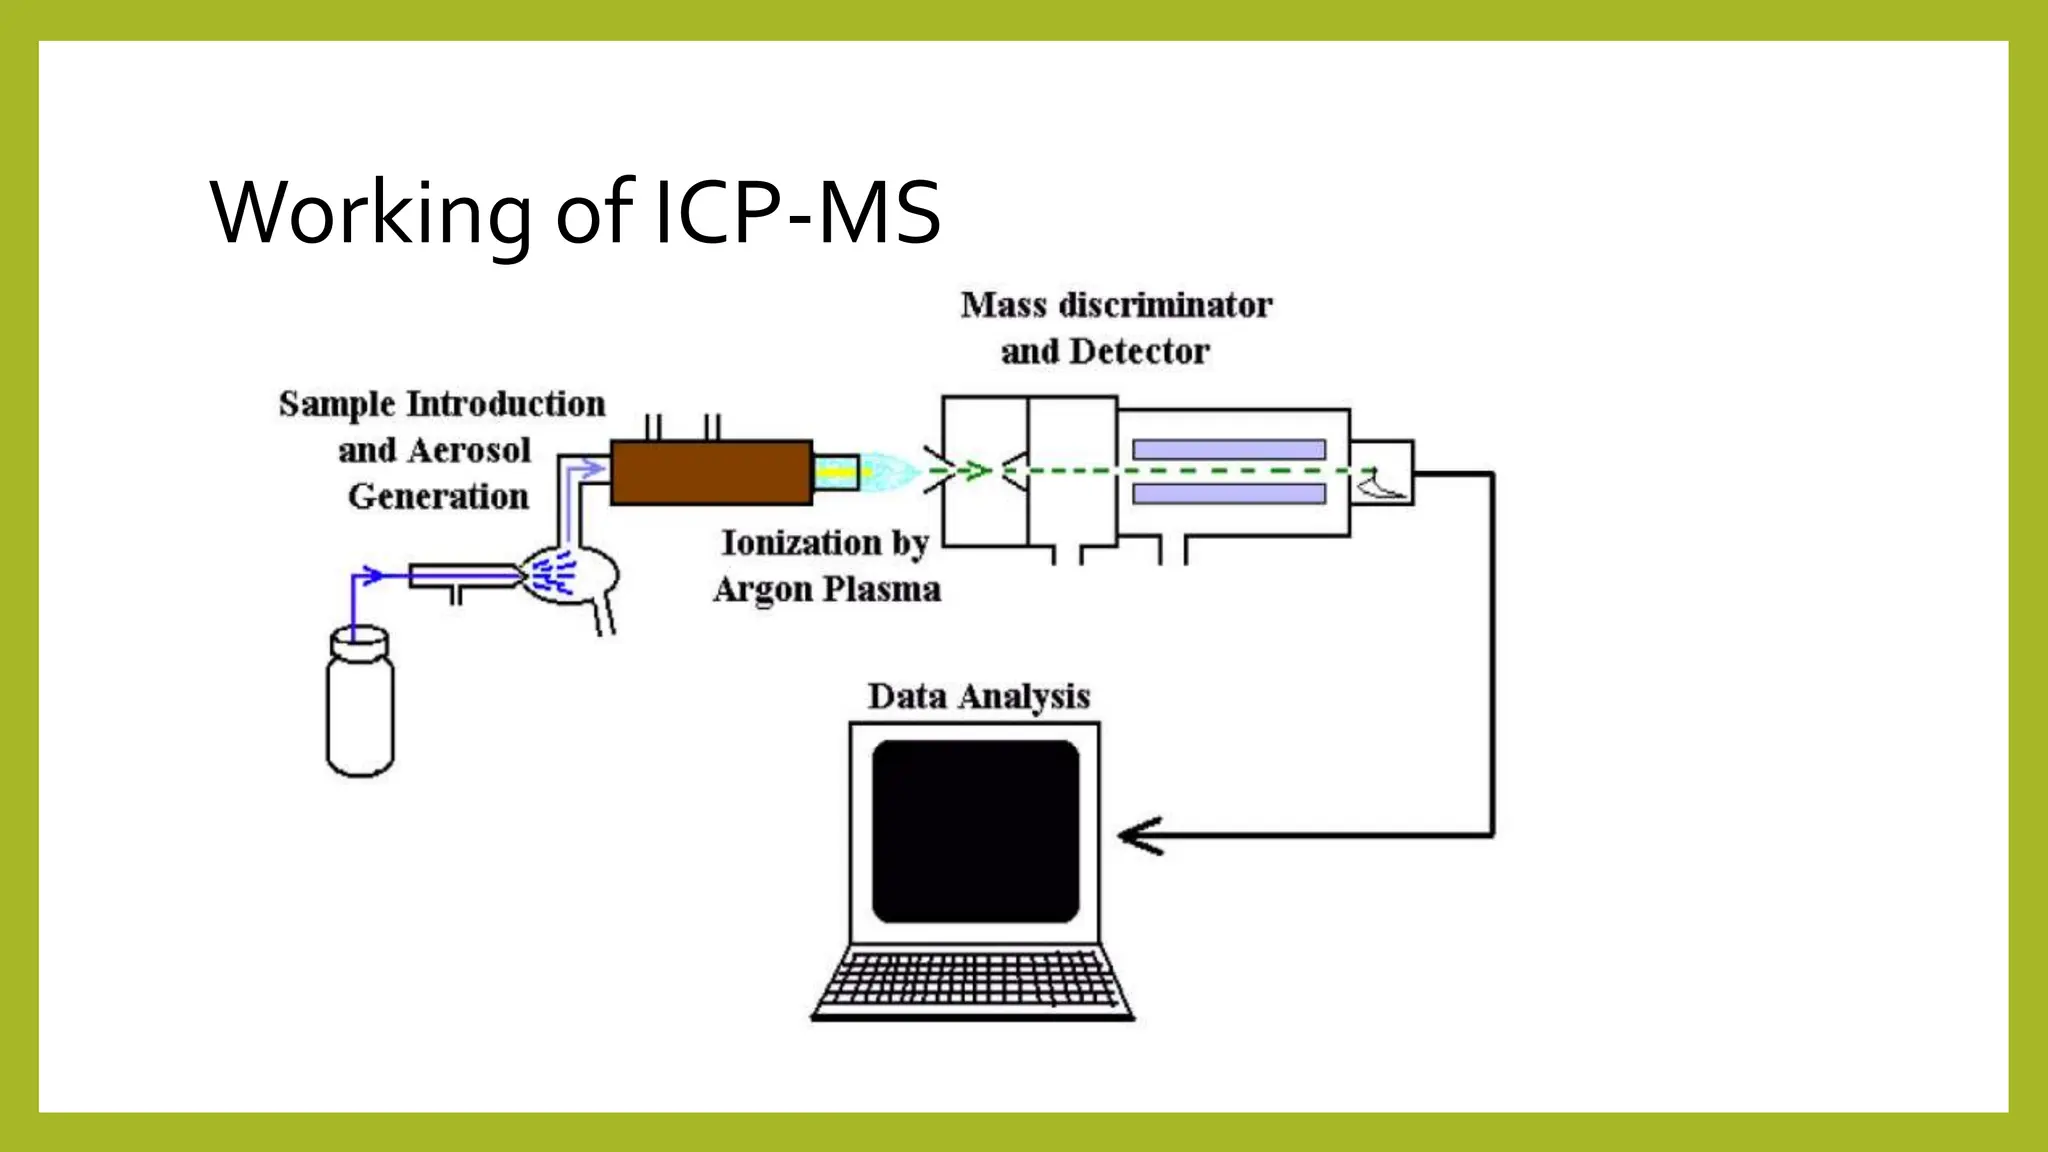
You take a liquid sample, turn it into a fine mist, push it into an argon plasma running at roughly 6000 to 10000 kelvin depending on the instrument, and the atoms and ions in that plasma emit light at wavelengths that correspond to their electron energy level transitions. The spectrometer separates that light and counts the photons. That's basically the whole thing. What makes or breaks the analysis is almost nothing to do with the theory and everything to do with sample introduction efficiency, spectral interferences, and whether your plasma is actually stable enough to give you consistent readings. I spent years running ICP-OES and ICP-MS, and the gap between textbook Principles Of Plasma Spectroscopy and what you're dealing with at 2 AM when a batch of environmental samples starts showing drifting baselines is enormous. Let me walk through the practical side of things.
Understanding The Core Principles Of Plasma Spectroscopy
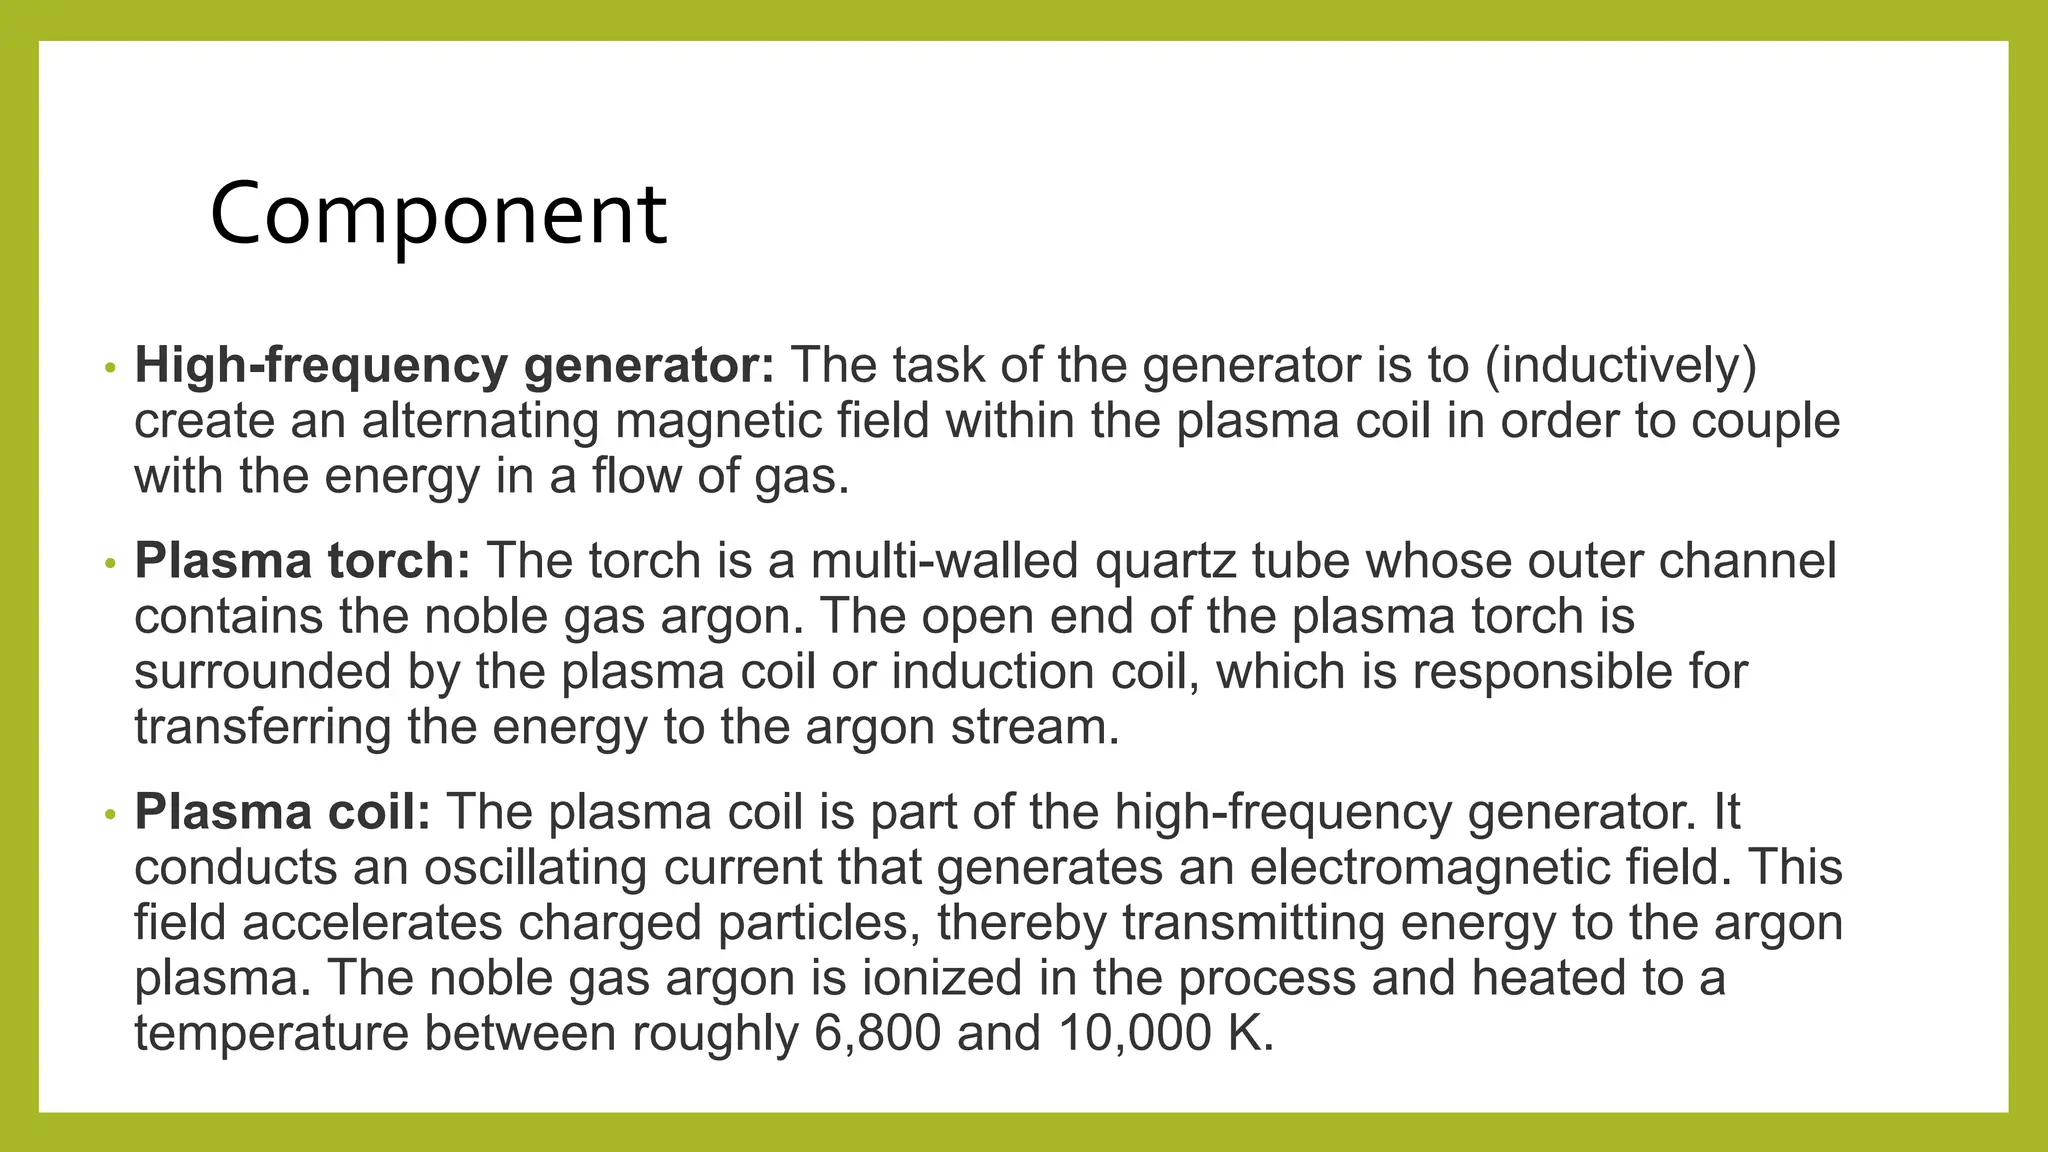
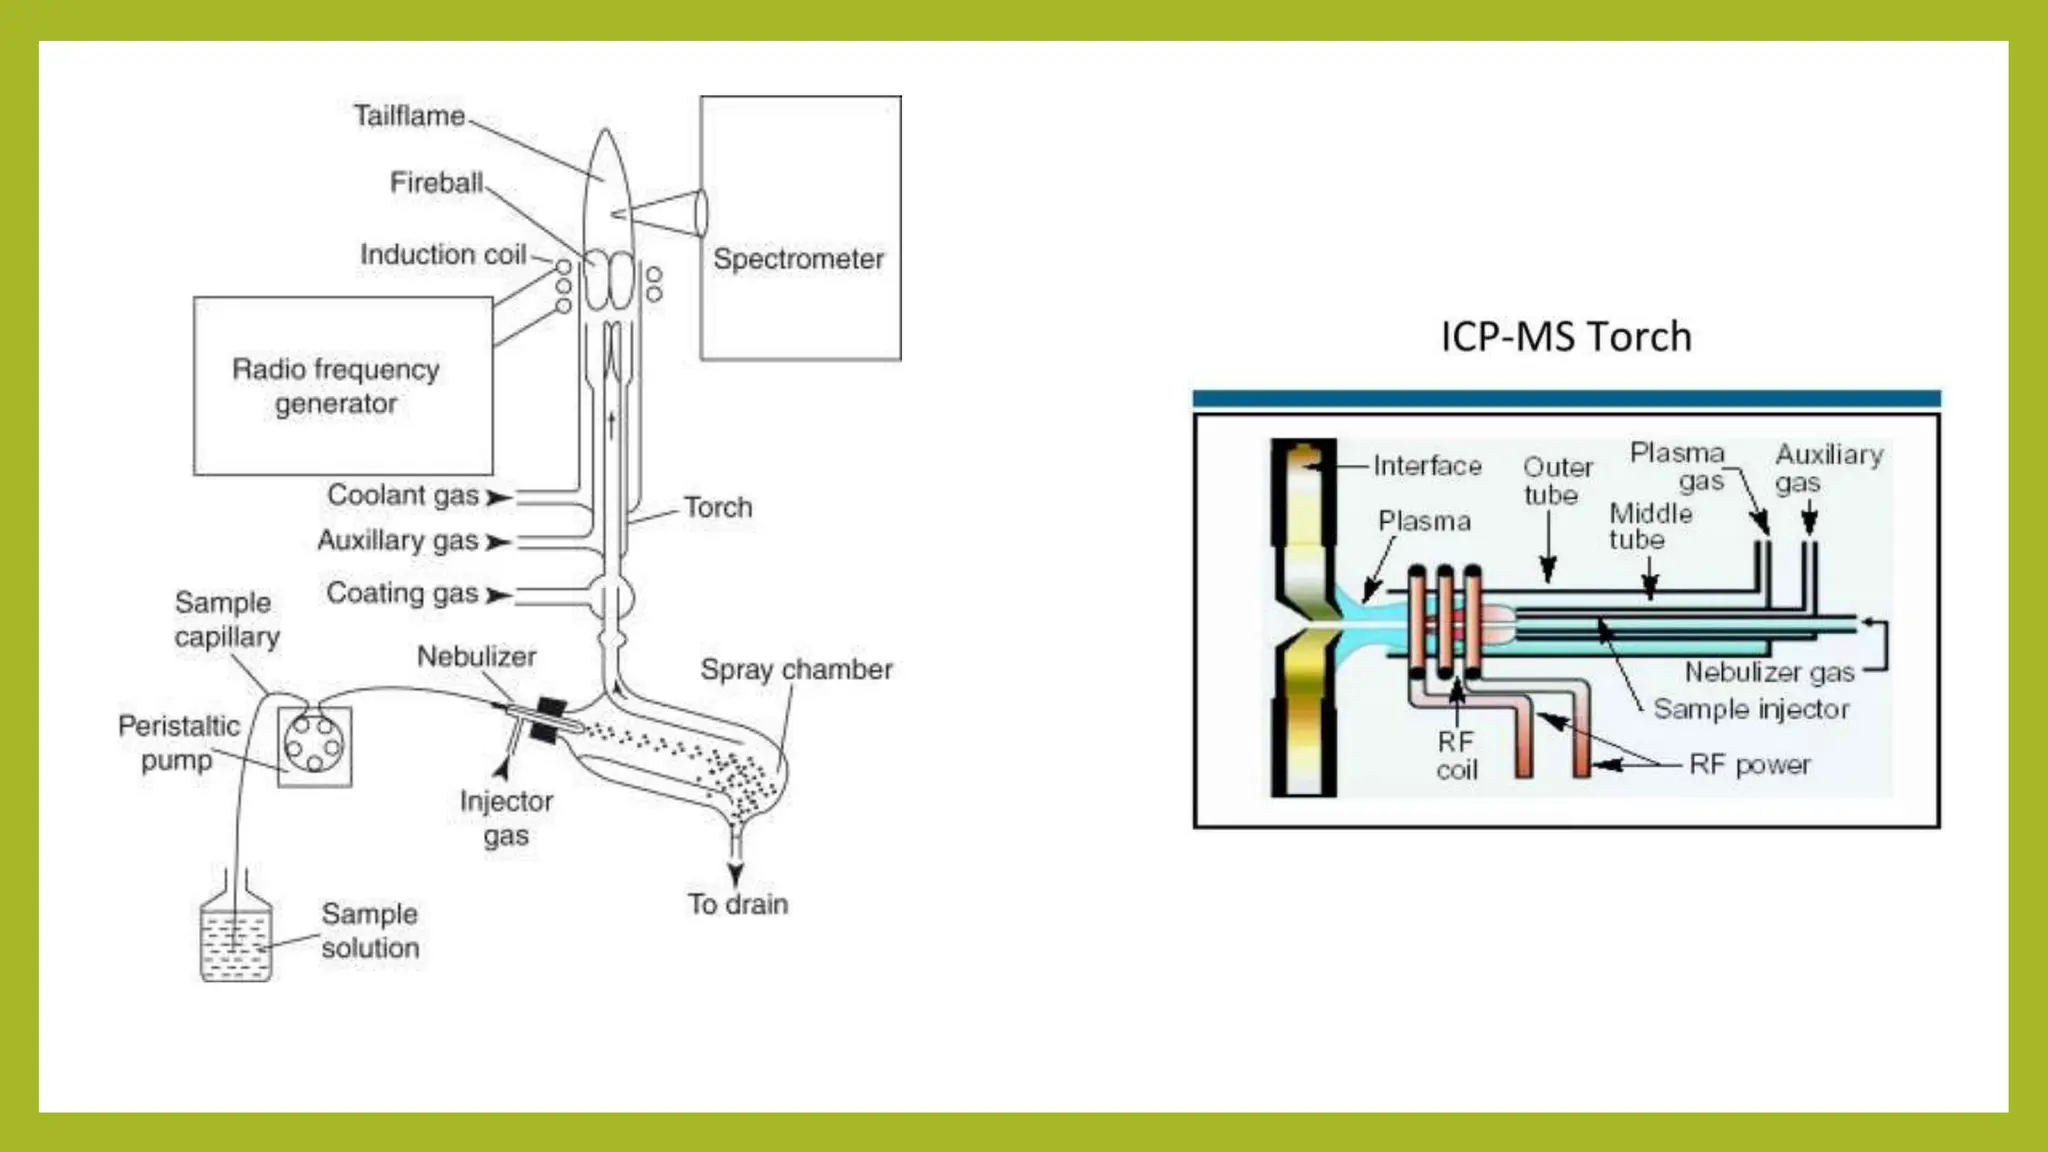
The basic mechanism involves three stages. First, the sample has to get into the plasma as a fine aerosol. Second, the plasma has to atomize and excite the analytes. Third, the emitted radiation has to be detected without too much background noise drowning out the signal. The sample introduction system is where most problems start. A cross-flow nebulizer paired with a Scott double-pass spray chamber is the standard setup. The nebulizer turns your liquid into droplets, most of which are too large to carry into the plasma. The spray chamber throws out the big droplets by impaction, and only the fine mist makes it through to the torch. If your nebulizer is partially clogged, which happens more often than you'd think with high-salinity samples, you'll see signal drop and stability deteriorate. I've replaced nebulizers that looked perfectly fine on the outside because a tiny bit of silica precipitation had narrowed the capillary internally. The plasma itself is sustained by induction coupling. Argon gas flows through a copper coil cooled by water, and a radio frequency generator at 27.12 or 40.68 MHz ionizes the gas to create the discharge. The torch usually has three concentric channels: the outer channel carries the plasma gas, the middle channel carries the auxiliary gas that lifts the plasma off the injector tube, and the inner channel carries the sample aerosol. Getting the auxiliary gas flow right matters. Turn it up too high and you push the plasma up and away from where the light collection optics are looking. Turn it down too low and you melt the injector tube. There's a narrow window and it shifts with age of the torch components and with changes in ambient conditions.
Once the sample is in the plasma, the heat strips electrons and excites the remaining ones. When those electrons fall back to lower energy states, they emit photons. The wavelength is element-specific, which is why this technique works for multi-element analysis. Intensity is roughly proportional to concentration, at least within a certain range, which is why you need calibration curves.
Get the Full Details
Where The Textbook Stops And Reality Begins
One thing that textbooks don't stress enough is that spectral interferences are a constant background problem. Line overlaps happen all the time, especially in complex matrices. I was analyzing seawater for trace metals and kept seeing elevated vanadium readings that didn't match my spike recovery. Turns out there's a Chromium-52 line sitting almost exactly on top of a Vanadium-51 line in many instruments. The spectral resolution of a mid-range CCD spectrometer wasn't enough to separate them cleanly. I had to switch to a different analytical line for vanadium and recalibrate everything. It added about 20 minutes per batch but saved me from publishing garbage data. Another thing nobody warns you about is matrix-induced signal suppression and enhancement. High total dissolved solids change the viscosity and surface tension of your sample, which changes nebulization efficiency. High sodium concentrations can depress ionization of other elements in the plasma. I once spent three days chasing a systematic under-recovery issue on lead in soil digests before realizing the calibration standards were in dilute nitric acid while the actual samples had significant amounts of dissolved aluminum and iron from the digest. The matrix matched standards fixed it immediately. For ICP-MS users, polyatomic interferences are their own special torment. Argon combines with oxygen, hydrogen, nitrogen, and chlorine in the plasma and sample matrix to form species like ArO+, ArCl+, and N2+ that sit on the same mass as your target analytes. Iron-56 is the classic casualty because ArO+ sits right on top of it. There are collision cell technologies and mathematical correction equations, but they're not foolproof. Sometimes you just need to acknowledge the limitation and move to an interference-free mass if your instrument allows it.
Calibration And Quality Control That Actually Work
External calibration with matrix-matched standards is the baseline approach and it works fine for simple matrices. For anything more complicated, standard addition is your safety net. You take aliquots of your sample, spike them with known increments of the analyte, and extrapolate back to find the original concentration. It's slow and it eats up sample, but it compensates for matrix effects that no amount of dilution will fully address. Your QC sample plan should include a certified reference material at the start and end of each run, a method blank, and at least one duplicate. If your CRM is off by more than two standard deviations from the certified value, the batch is suspect. Don't second-guess that. I've seen labs push forward with biased batches because someone didn't want to redo the work. That's how bad data gets published. Internal standards are non-negotiable for ICP work. You add a known concentration of an element that isn't present in your samples to every standard, blank, and unknown. Scandium, yttrium, rhodium, and indium are common choices depending on your mass range. The internal standard corrects for drift in nebulization efficiency, plasma conditions, and instrument sensitivity over time. If your internal standard signal is drifting by more than five percent, something is wrong and you should investigate before trusting any data from that run.
Practical Troubleshooting
If your signals are slowly drifting down over a run, the most likely culprits are a clogged nebulizer, a worn spray chamber tip, or contamination buildup on the torch injector. Clean or replace as needed. I keep a spare injector tube on hand and swap it every few weeks preemptively rather than waiting for failure during a production run. If your background is elevated across the spectrum, check your argon purity. Low-grade argon with high oxygen and moisture content will increase the continuous background and can also oxidize components inside the torch assembly over time. Ultra-high purity argon, 99.996 percent or better, is worth the cost. If you're getting inconsistent replicate measurements, look at your sample introduction first. Bubbles in the tubing, a loose connection, or an air leak at the nebulizer will cause exactly this kind of noise. Run the carrier line with just acid and watch the signal stability before introducing samples. Base RSD under one percent on a mid-range standard is a reasonable target for a well-tuned system.
Detection limits vary by element and by instrument configuration. For most elements in ICP-OES, you're looking at low parts per billion in solution. ICP-MS pushes that into the parts per trillion range for many elements. But detection limit is not the same as practical quantification limit. Your actual limit depends on your matrix, your cleanup procedure, and how clean your reagents are. Ultra-pure nitric acid and deionized water at 18 megohm centimeters or better are minimum requirements. Anything less and your blanks will eat your detection limits alive. The instruments themselves have gotten far more robust over the last decade. Automated wash cycles, self-diagnostic routines, and better software corrections have reduced the daily maintenance burden significantly. But the underlying physics hasn't changed. You still need to understand what's happening in that plasma, respect the limitations of your detection system, and validate your results against known materials. The Principles Of Plasma Spectroscopy are straightforward. Applying them correctly is where the experience counts.