Why Your Recombinant Protein Keeps Precipitating Out of Solution
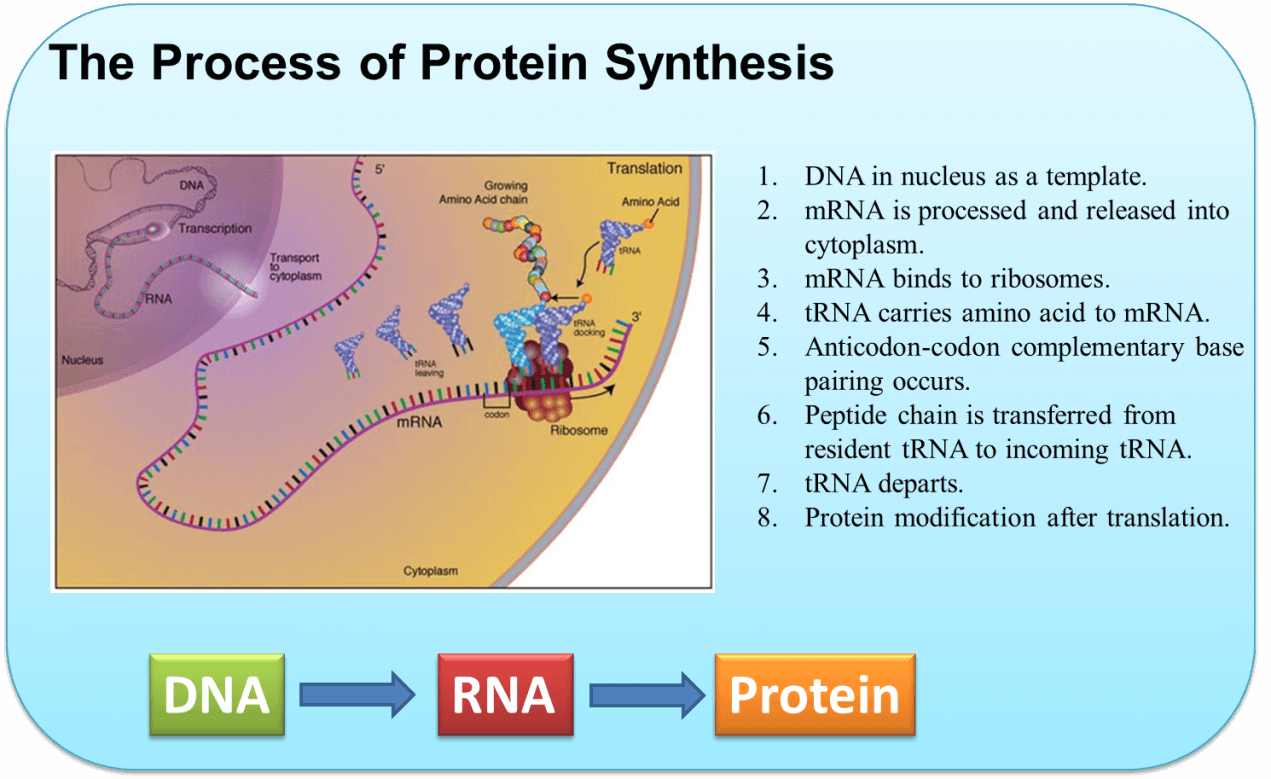
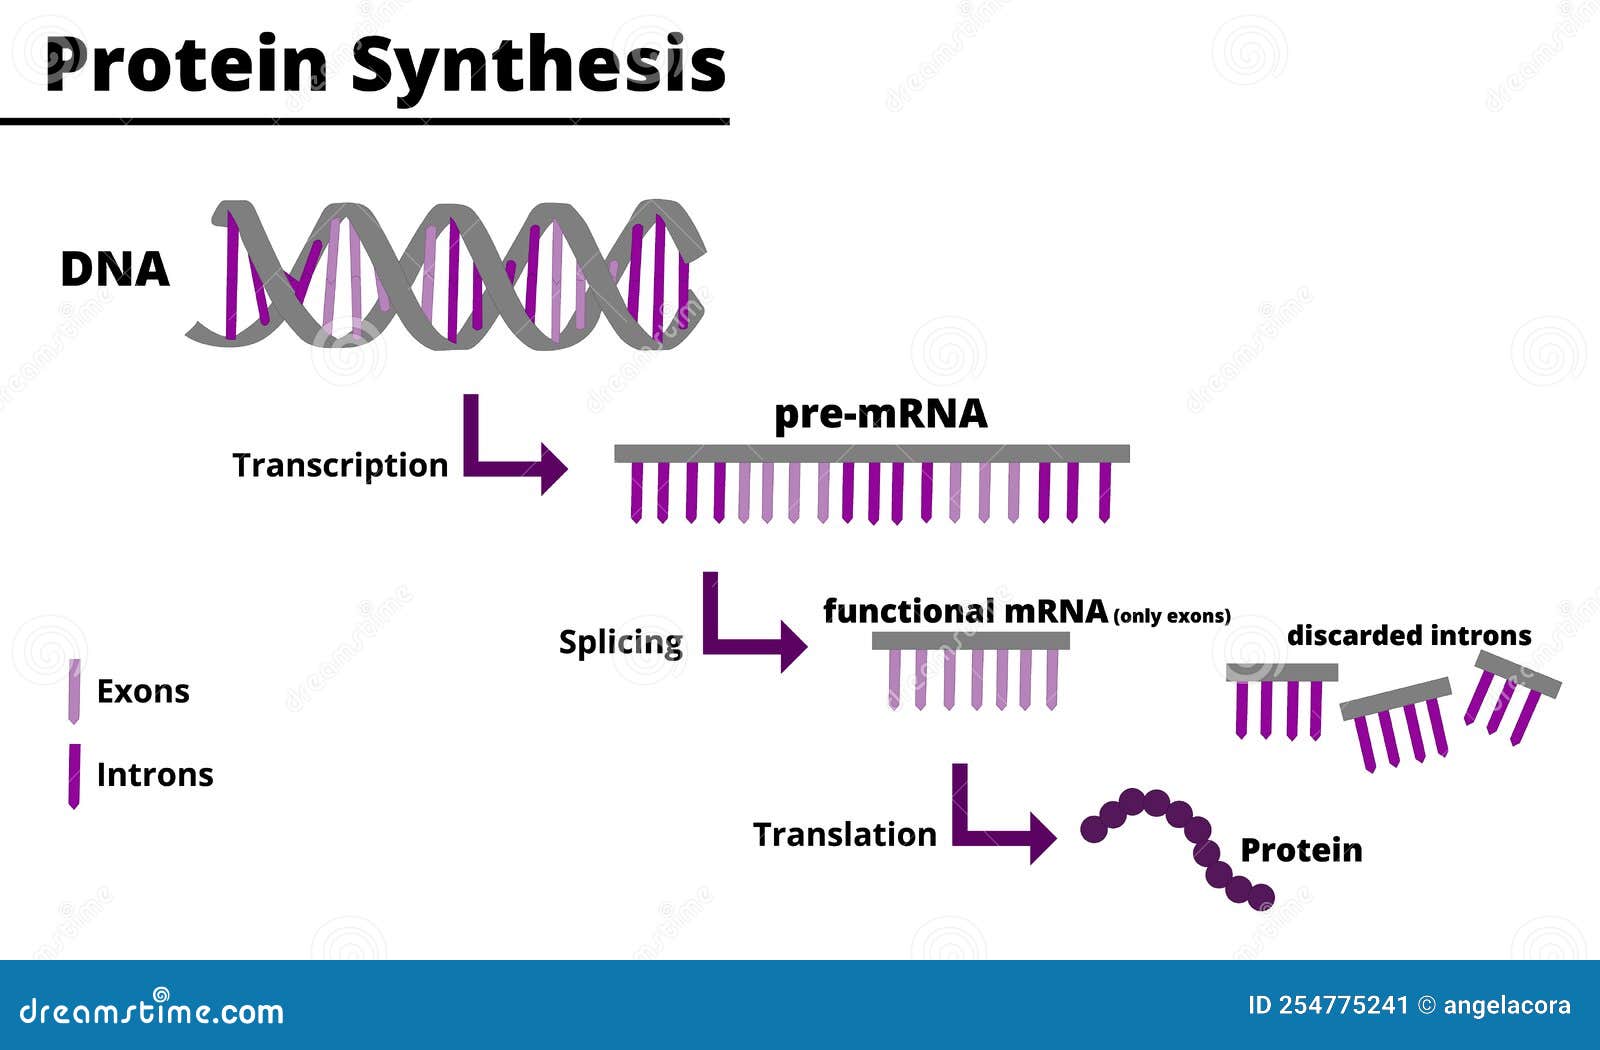
You've probably seen this before. You optimize the culture conditions, you induce expression at the right OD, you harvest and lyse, and then you're left with either inclusion bodies or a supernatant that looks clear but contains zero functional protein when you run your western blot. The issue isn't usually the protocol itself. It's understanding what actually happens during the Process Of Protein Formation and why it falls apart under certain conditions. Protein formation happens in two main stages. First there's transcription, where DNA gets copied into messenger RNA by RNA polymerase. Then translation kicks in, where ribosomes read that mRNA and string amino acids together into a polypeptide chain. The chain doesn't fold immediately. It emerges from the ribosome and goes through co-translational folding, post-translational modifications, and eventually reaches its native conformation. In eukaryotes this involves the endoplasmic reticulum and Golgi apparatus. In bacteria, which is what most people use for recombinant work, it's messier and happens in the cytoplasm without those organelles.
The Process Of Protein Formation in Practice
When you're actually doing this in a lab, the real bottleneck isn't getting the gene expressed. It's getting it to fold correctly. I spent about six months working with a membrane protein that refused to stay soluble no matter what I changed. Temperature, expression strain, inducer concentration, chaperone co-expression - I tried everything from 16 degrees Celsius up to 37, tried BL21(DE3), Rosetta, Origami, SHuffle. The protein would express fine at the mRNA and initial polypeptide level, but it kept aggregating into inclusion bodies every single time. The workaround came from understanding that the problem wasn't expression level. It was the rate of translation. When you induce at high temperature with strong promoters like T7, the ribosomes move fast enough that the protein doesn't have time to fold properly as it comes off the ribosome. I dropped the induction temperature to 18 degrees, cut the IPTG concentration to 0.1 millimolar, and extended the induction time to 16 hours instead of the usual 4. That gave the folding machinery enough time to work. Soluble yield went from essentially zero to about 8 milligrams per liter of culture. It was that simple once I stopped treating it like an expression problem and started treating it like a folding problem. There are some things beginners consistently get wrong about protein formation. The first is assuming more inducer equals more protein. That's almost never true for folded, functional protein. Higher inducer concentrations push the ribosomes harder, which means faster translation and more misfolding. The second mistake is thinking that soluble protein is always folded protein. A cleared lysate can contain a lot of misfolded or partially folded protein that just happens to stay in solution. You need functional assays, not just SDS-PAGE, to confirm you actually have the right product.
Another counter-intuitive point is that codon optimization isn't always better. I had a case where fully optimizing the codons for E. coli actually made the problem worse. The optimized gene was translated so efficiently that the ribosomes crowded the mRNA and the protein couldn't fold between successive rounds of synthesis. We back-substituted a moderate number of rare codons at strategic positions, which slowed down translation at key points and gave the protein time to fold domain by domain. Yield improved despite the gene being "less optimized." The Process Of Protein Formation also breaks down completely in certain scenarios. If your protein has disulfide bonds and you're expressing it in the reducing environment of the E. coli cytoplasm, it won't form properly regardless of what you do. You'd need to target it to the periplasm using a signal sequence like pelB, or use oxidizing strains like SHuffle or disulfide bond helper variants. Even then, not all proteins fold correctly in the periplasm. Some need chaperones, some need specific post-translational modifications that bacteria can't do, like glycosylation. If you need glycosylation, bacterial expression is the wrong system and you should move to yeast, insect cells, or mammalian culture upfront instead of wasting weeks on something that will never work. Purification is where a lot of the damage happens too. His-tag purification is convenient but it's not gentle. Imidazole concentrations, low pH elution, and repeated freeze-thaw cycles can denature sensitive proteins. I learned this the hard way with a kinase that lost activity after five purification cycles even though it looked pure on the gel. Switching to a tobacco etch virus cleavage site and running a size exclusion chromatography step after the affinity step fixed the activity problem. SEC also removes aggregates that you wouldn't see on a standard gel.
Get the Full Details
If you're working with difficult proteins and the bacterial system keeps failing, consider trying expression at lower temperatures for longer times before moving to more expensive systems. A 16-degree overnight induction with 0.1 mM IPTG costs nothing and takes maybe an extra day. Switching to Pichia or baculovirus systems runs into hundreds of dollars per construct and still might not work. Try the cheap fixes first. The fundamental takeaway is that protein formation isn't just about making the polypeptide chain. It's about giving that chain the time and conditions it needs to reach its correct three-dimensional structure. Expression is the easy part. Everything after that is where the actual work happens.