Why Your Chemoselectivity Keeps Failing
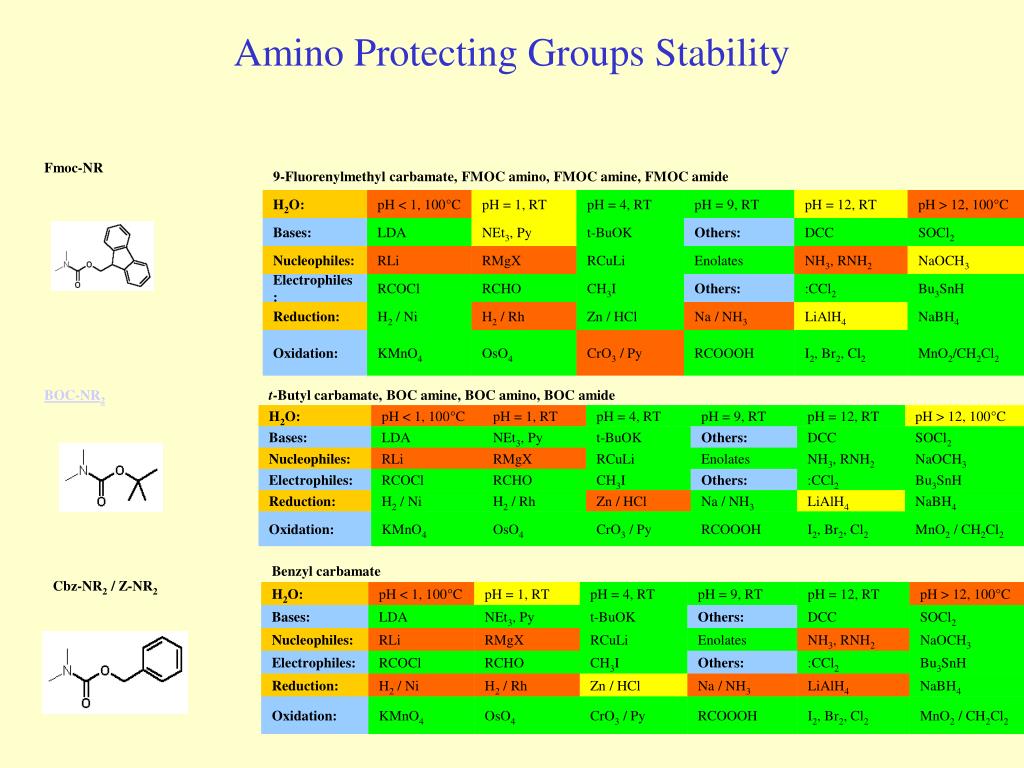
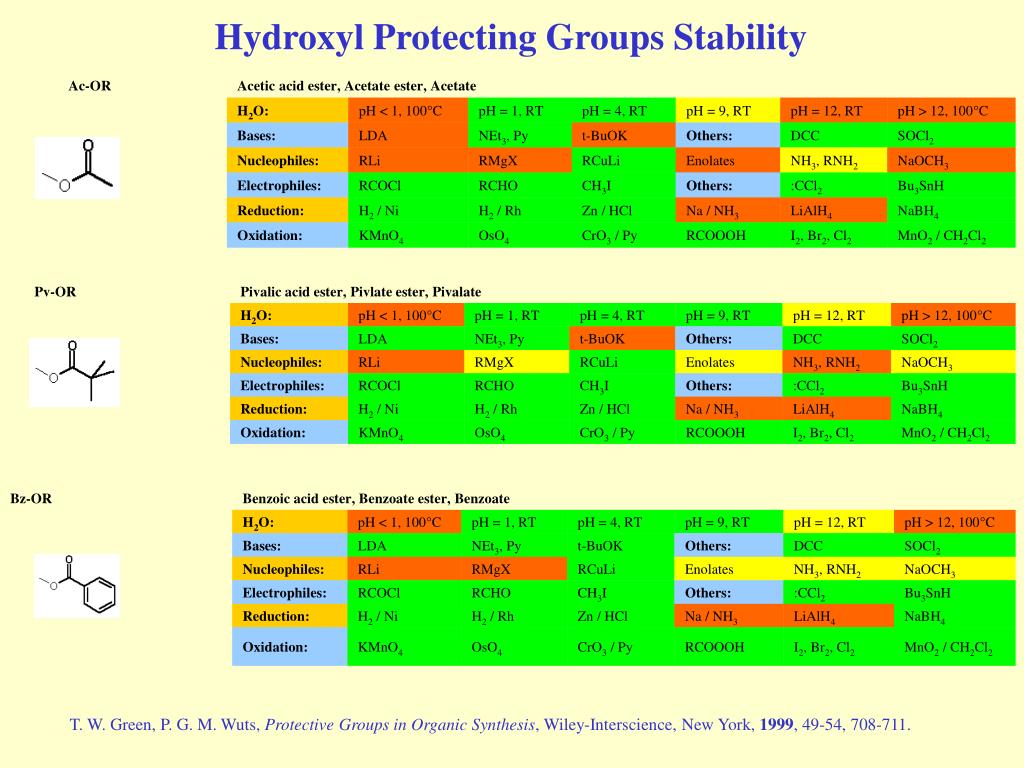
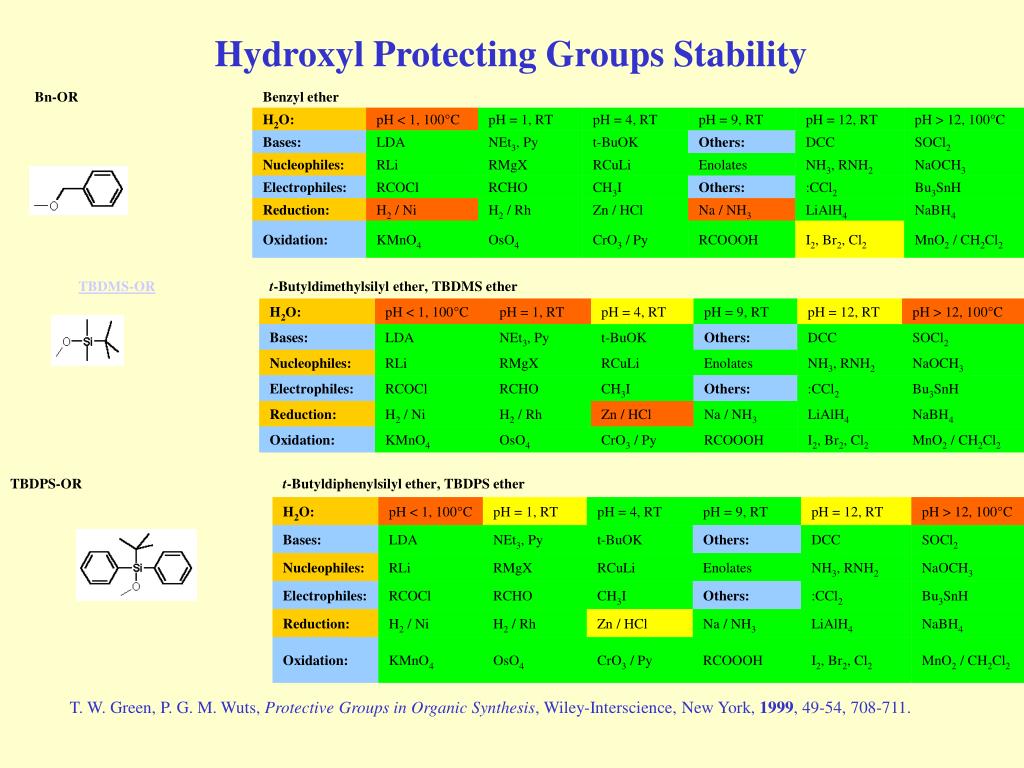
Protecting groups exist because molecules rarely behave the way you want them to when you introduce a reagent. If your target molecule contains both a free hydroxyl and a free amine, and you want to acylate only the oxygen, standard conditions will attack the nitrogen first. Period. There is no workaround except to mask the amine before you proceed, remove it later, and move forward. This is the fundamental reason protective groups are baked into almost every multi-step synthesis. The groups I reach for most often depend entirely on what functionality needs temporary suppression. For alcohols, TBDMS (tert-butyldimethylsilyl) and TBDPS (tert-butyldiphenylsilyl) ethers dominate routine work. TBDMS installs quickly with TBDMSCl and imidazole in DMF, and removes with TBAF in THF within thirty minutes at room temperature. TBDPS is bulkier and significantly more stable to acidic conditions, which makes it useful when you plan to carry the alcohol through multiple sequences before final removal. Amines are almost always masked as carbamates. BOC (tert-butoxycarbonyl) is removed with TFA in DCM, typically quantitative within ten to fifteen minutes. Cbz (benzyloxycarbonyl) comes off with hydrogenolysis over Pd/C or with HBr in acetic acid. Fmoc is the only one that drops under basic conditions — 20 percent piperidine in DMF, usually complete within twenty minutes. Each one has a different removal profile, and mixing them up in a single synthesis without tracking which is which is a fast route to a very unhappy morning.
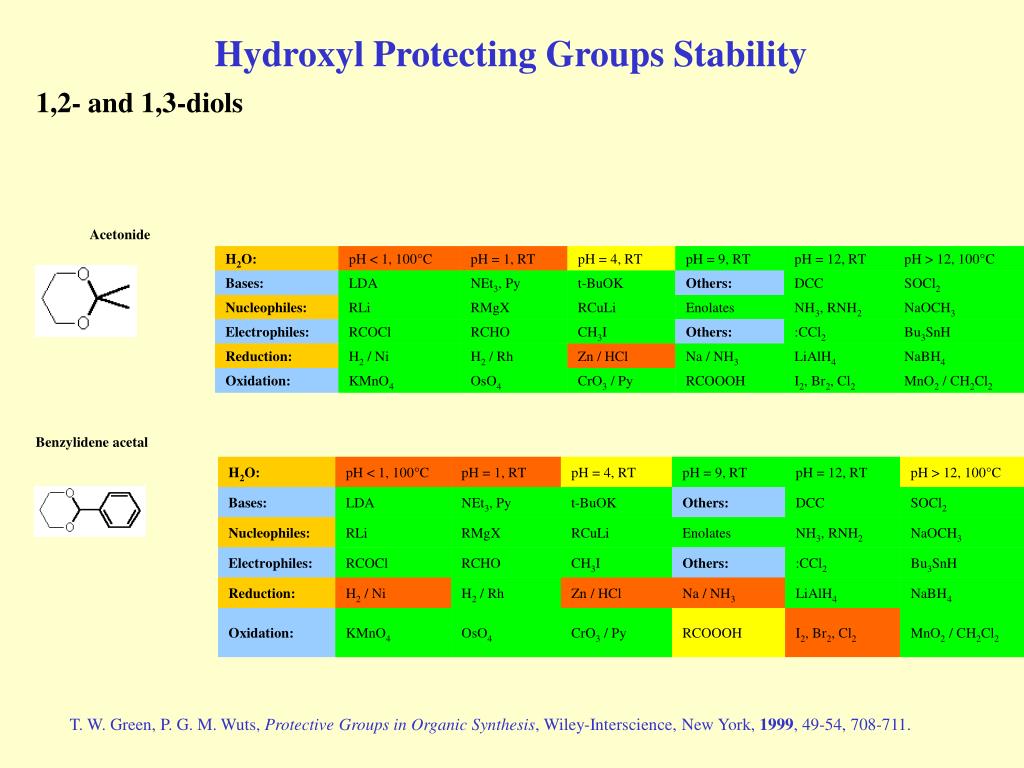
Carbonyls are trickier. Dimethyl acetals and dimethyl ketals protect aldehydes and ketones under acidic conditions with a Dean-Stark trap or molecular sieves. They come off with aqueous acid. Dithianes require a sulfur to carbon bond formation using 1,3-propanedithiol and BF3·OEt2, and they remove with heavy metals like HgCl2 or I2 in aqueous MeOH. This is slow and produces sulfur-smelling waste, but dithianes survive strong bases and nucleophiles that would destroy an acetal.
How the Removal Actually Works in Practice
Fluoride-mediated silyl deprotection is not as simple as the textbooks make it look. TBAF in THF does cleave TBDMS ethers rapidly, but it also promotes elimination adjacent to secondary and tertiary alcohols, and it can open epoxides or attack -keto esters. I once had a substrate with a -keto ester three atoms away from a TBDMS-protected hydroxyl. The TBAF didn't just strip the silyl group. It deprotonated the active methylene, and the resulting enolate triggered an intramolecular elimination that destroyed the product. I switched to acetic acid in methanol, which is milder and slower — about four hours at room temperature for complete deprotection — but it left the ketone untouched and gave clean recovery of the free alcohol. BOC removal with TFA is straightforward until it isn't. TFA will cleave Boc from a standard secondary amine in minutes. But if your molecule contains an acid-labile protecting group elsewhere — say an acetal or a trityl ether — the same TFA solution that removes the BOC will also start attacking that group. I've seen trityl leave almost as fast as BOC under identical conditions, which turns a selective deprotection into a messy mixture you have to sort out by chromatography the hard way. The fix is usually to use milder acid like 4M HCl in dioxane for the BOC removal, which takes longer — roughly forty-five minutes to an hour — but spares the trityl. Hydrogenolysis of Cbz is reliable on small scale but becomes unpredictable at larger scale. The reaction works fine with 10 percent Pd/C under a balloon of H2 in MeOH, but at half a mole scale, heat management becomes an issue. The exotherm can push the temperature up enough to cause hydrogenolysis of other sensitive groups or promote unwanted reductions of alkenes you intended to keep intact. I usually cool the reaction to 0 degrees Celsius and add the Pd/C in small portions over twenty minutes rather than dumping it all in at once. It slows the reaction but keeps it controlled.
Get the Full Details
The Orthogonality Trap Most People Walk Into
The most common mistake I see in retrosynthetic plans is assuming that two different protecting groups are fully orthogonal when they are not. BOC and Cbz seem orthogonal — one comes off with acid, the other with hydrogenolysis. That is true in isolation. But if your final step requires Pd/C hydrogenation and your molecule contains a benzyl ether or a styrenyl double bond, you just lost selectivity. The Pd/C will reduce the alkene and cleave the benzyl ether alongside the Cbz. I've corrected this kind of error after a colleague spent three days on a deprotection sequence only to realize the benzyl protecting group they thought was safe was gone. Another overlooked interaction is residual TBAF from a prior silyl deprotection step. Even after aqueous workup, trace fluoride can remain bound to silicate species in the organic layer. If you then attempt an acid-catalyzed step with your crude material, that residual fluoride can catalyze side reactions that don't appear if you purify the intermediate first. Loading the NMR shows you what is actually there. Don't skip it.
When Protecting Groups Fail Completely
Sterically congested silyl ethers sometimes refuse to install cleanly. TBDMS chloride with imidazole in DMF works beautifully for primary and secondary alcohols. For a hindered secondary alcohol next to a quaternary center, you may need to switch to 2-chlorotrimethylsilane with a stronger base like DBU, or abandon the silyl route entirely and use an ether instead. Benzyl ethers require NaH and BnBr in DMF, which is a harsh combination. If your substrate contains an ester or a lactone, the base will hydrolyze or open it before the protection finishes. Acetal formation from aldehydes is equilibrium-limited. You drive it forward by removing water, but if your aldehyde is prone to polymerization or self-condensation under the acidic conditions required for acetal formation, you will lose material before the protected product forms in good yield. I once worked with an aromatic aldehyde bearing a free phenol. The acidic conditions caused the phenol to interfere with the acetal equilibrium through hydrogen bonding, slowing the reaction dramatically. Switching to a thioacetal using 1,3-propanedithiol and a Lewis acid gave quantitative conversion in two hours. Fmoc removal with piperidine generates dibenzofulvene as a byproduct, which can alkylate nucleophilic sites on your substrate if it isn't scavenged. Adding a small amount of phenol or N-methylmorpholine to the piperidine solution traps the fulvene and prevents secondary reactions. Without this, you sometimes see unexpected N-alkylated byproducts that are difficult to separate from your desired product.
A Note on Scale and Practicality
Protecting group strategies add steps to a synthesis, and each step costs time, reagents, and purification. A single protection-deprotection pair typically adds two operations and a chromatography or crystallization. On a multigram scale, that translates to measurable loss of material. I usually try to minimize the total number of protecting groups installed in any sequence. If I can achieve selectivity through reagent choice or temperature control rather than an extra protection step, I take it. Protecting groups are a tool, not a default answer. The chemistry behind Protective Groups In Organic Synthesis is well established, but the practical execution depends on knowing which removal condition will selectively affect only the group you intend to remove. That knowledge comes from running these reactions repeatedly and learning where the exceptions live. Every substrate behaves slightly differently, and the literature conditions are a starting point, not a guarantee.