Getting Real Work Out of Qualitative Analysis in the Lab
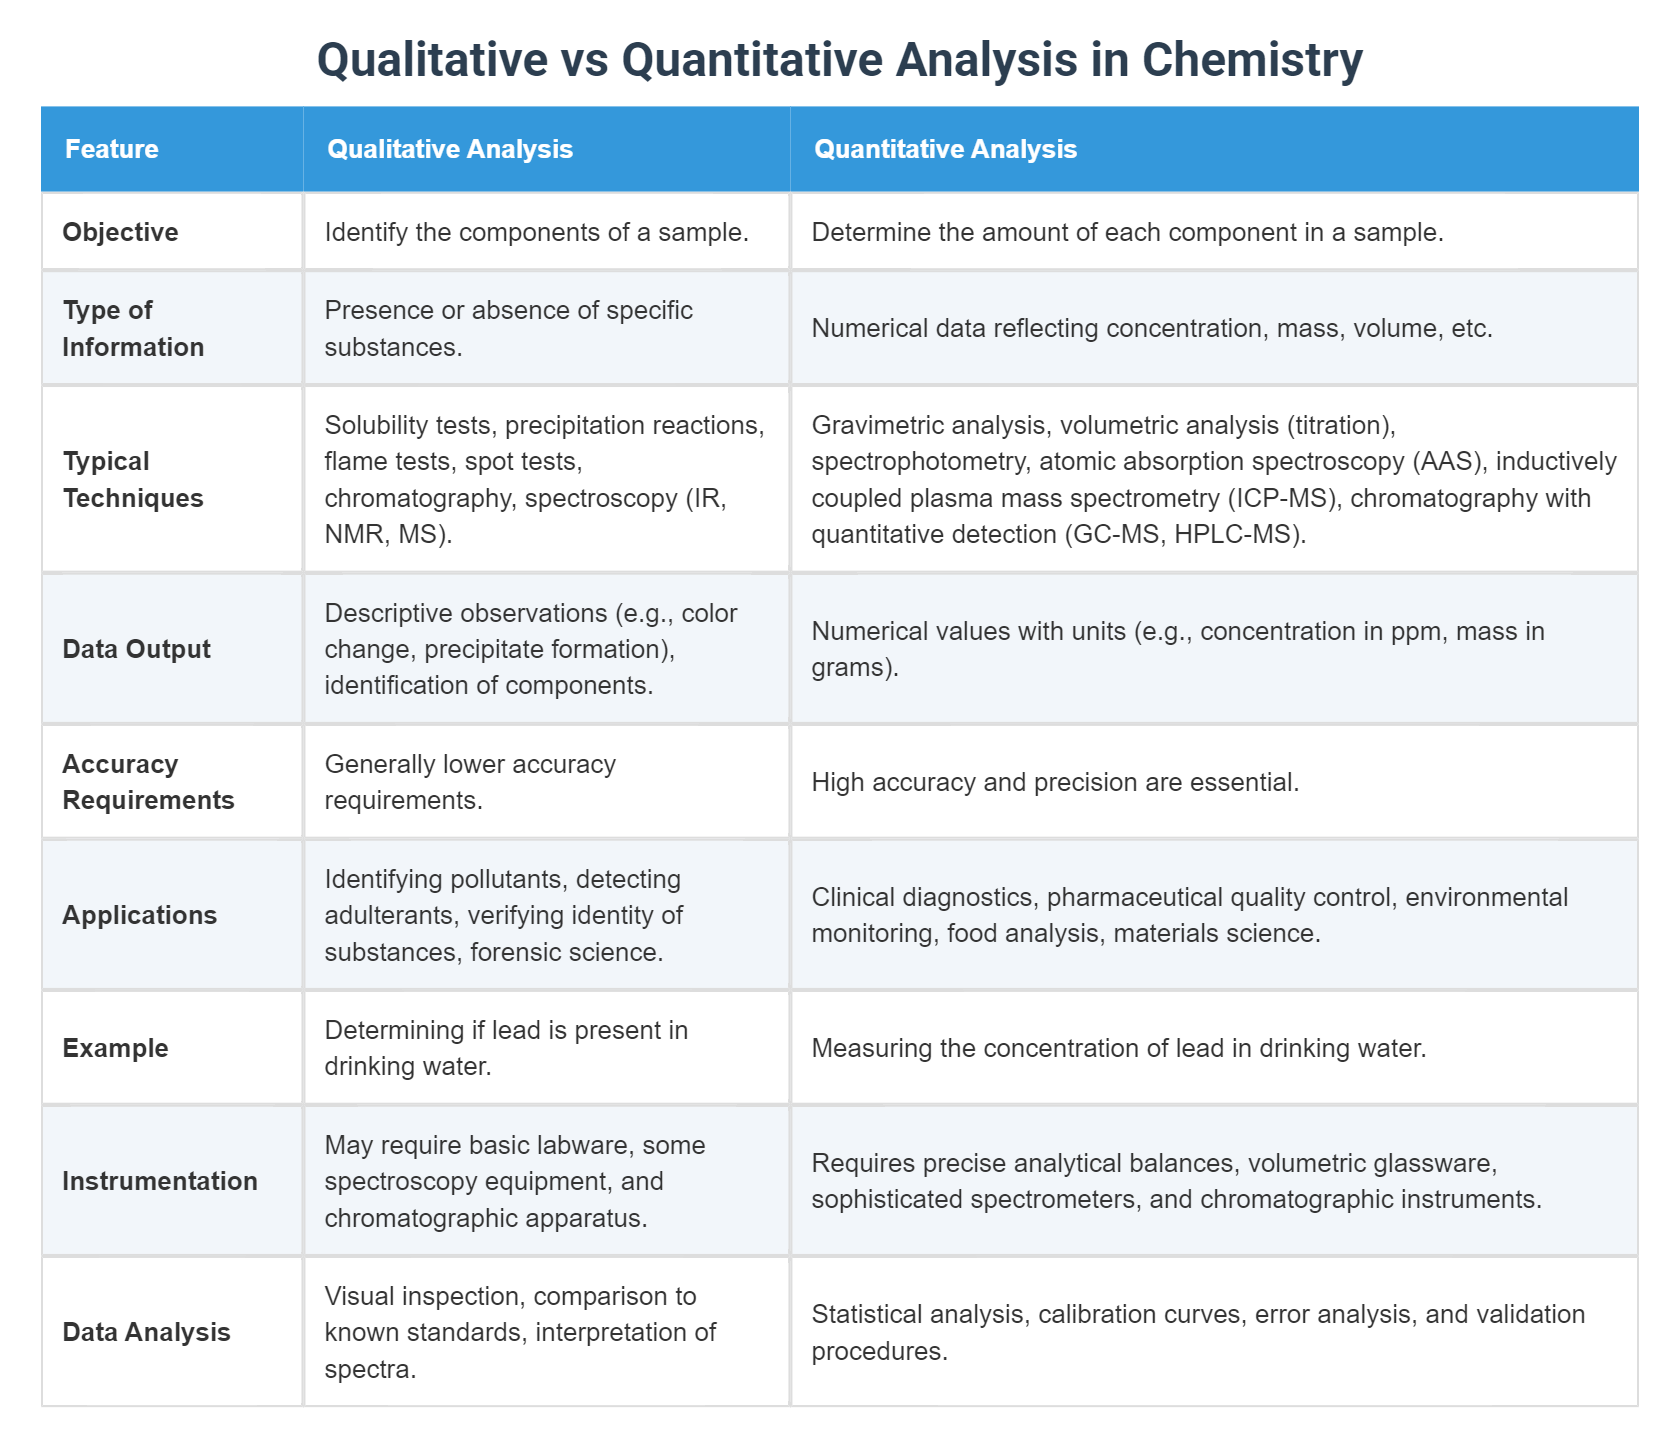
Qualitative analysis in chemistry is about figuring out what's in something, not how much. You add reagents to unknown solutions, watch for precipitates, color changes, gas evolution, and use that chain of observations to identify cations and anions present. It sounds simple on paper. The lab bench reality involves a lot more variables than the flowchart suggests. I spent years running these labs with undergraduates and dealing with samples that refused to behave the way textbooks promised they would. Here's what actually happens when you try to do this work properly.
Qualitative Analysis Chemistry Lab Workflow
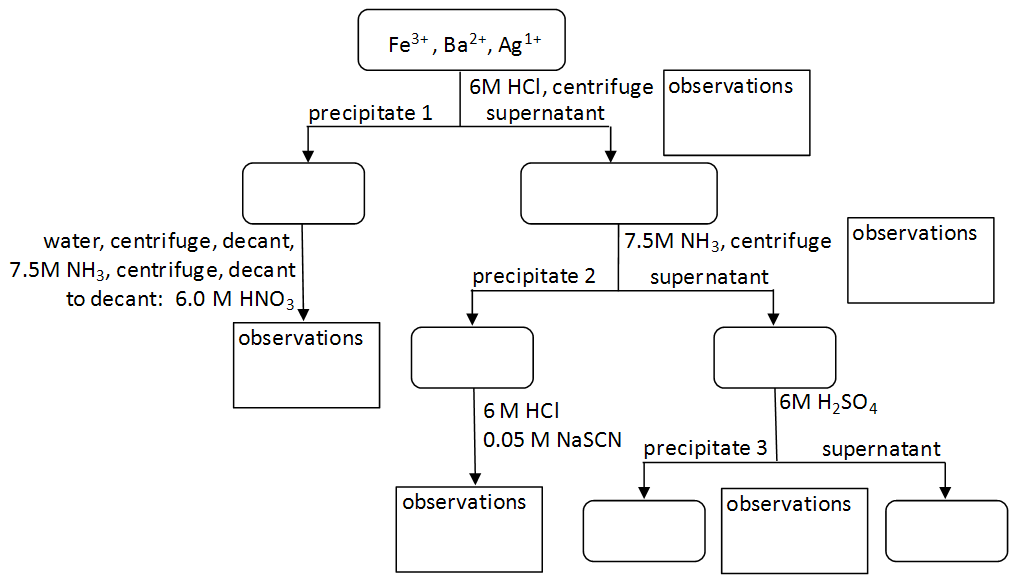
The standard approach starts with group separation. You take your unknown and systematically precipitate ions in groups using specific reagents. Group 1 comes down as chlorides with dilute HCl. Group 2 as sulfides in acidic medium with H2S or thioacetamide. Group 3 as hydroxides or sulfides in basic ammonia buffer. Groups 4 and 5 follow with carbonate and soluble salt separations respectively. Each group then gets individual confirmation tests. Here's the thing nobody emphasizes enough: the pH has to be correct at every single step. If your ammonia-ammonium chloride buffer for Group 3 is off by even 0.5 pH units, you'll either co-precipitate Group 4 ions or miss them entirely. I once had a student spend forty-five minutes convinced their unknown contained no Group 3 cations at all, only to discover the buffer had absorbed enough atmospheric CO2 during preparation that the pH had dropped to about 8 instead of the required 9 to 9.5. The iron and aluminum were still there, just not precipitating. We remade the buffer and got the results in ten minutes. Another thing that matters more than students think: the order of adding reagents. When you're testing for silver in Group 1, you acidify first with nitric acid, then add HCl. If you add the HCl before acidifying, you might accidentally precipitate something like lead chloride alongside the silver and barium chlorides, which would throw off your entire group analysis. The sequence is not a suggestion. It's the difference between a clean separation and a muddied result that takes three extra hours to untangle.
What Actually Goes Wrong and How to Fix It
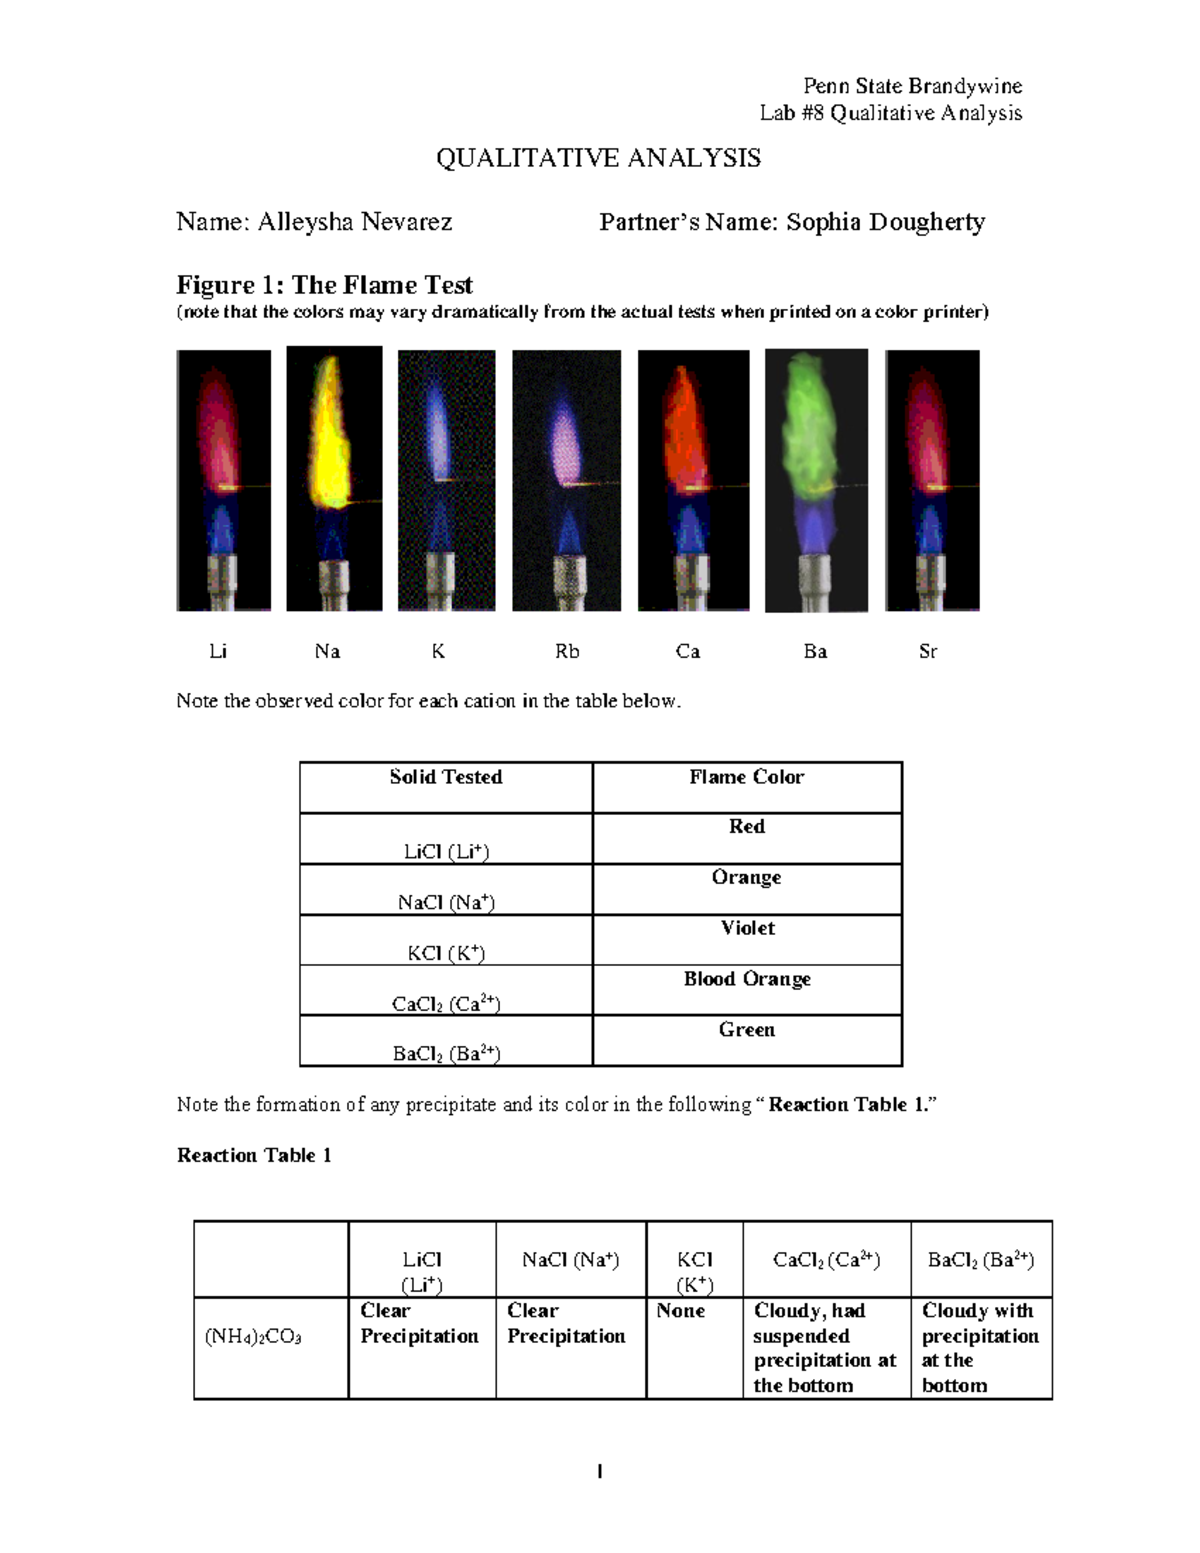
Precipitates are notorious for going supersaturated. You add the right reagent, wait the recommended two minutes, and nothing happens. The ion is absolutely there. You can confirm it by scratching the inside of the test tube with a glass rod to provide nucleation sites, or by cooling the tube in an ice bath. I've seen people discard valid samples because they didn't give supersaturated solutions the benefit of the doubt. Confirmatory tests are where most students lose confidence. The flame test for sodium is practically universal contamination — if your wire loop wasn't cleaned properly in hydrochloric acid, you'll see yellow every time regardless of what's in the sample. Clean it by dipping in concentrated HCl and holding it in a non-luminous Bunsen flame until no color appears. Then test again. Same thing with potassium permanganate used to test for reducing agents. If your glassware has any organic residue from previous labs, you'll get false positives regularly. Acid wash and rinse with distilled water, not tap water. Sulfide precipitates can oxidize over time, turning from their characteristic colors into brownish messes that look nothing like the textbook descriptions. Black PbS turning brown, yellow CdS developing a dull surface. If you're not reading results within fifteen to twenty minutes of precipitation, document what you see immediately and move on. Don't trust your memory against an oxidizing precipitate.
Get the Full Details

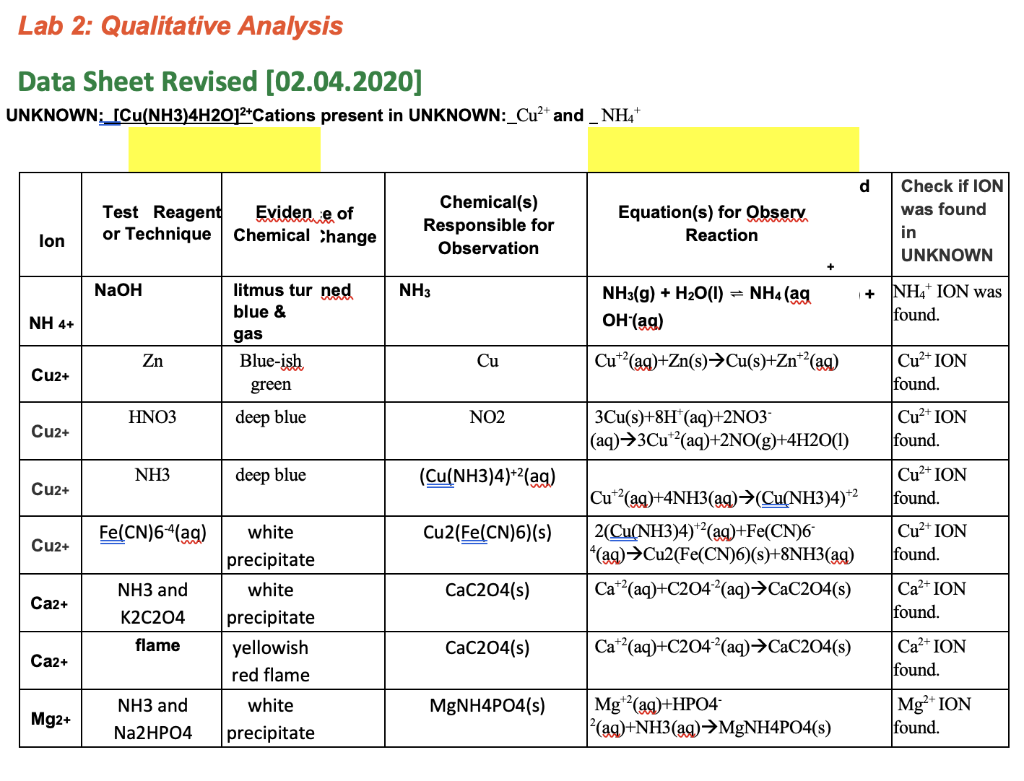
Ammonia complex formation is another area where assumptions fail. You add excess ammonia to a precipitate expecting it to redissolve, confirming a specific ion, but sometimes the precipitate simply doesn't dissolve fast enough at room temperature. Warm it slightly. A warm water bath for a minute or two is standard practice, not a deviation from procedure. Cold solutions react slower and less completely, and students often interpret the lack of dissolution as a negative result when it's just kinetic.
Limitations You Need to Accept
Classical qualitative analysis has real constraints. It typically requires concentrations in the range of 0.01 to 0.1 molar. If your unknown is diluted too far, you won't see precipitation or color changes even when the target ion is present. I've encountered samples where the instructor had accidentally prepared the unknown at 0.001 M and half the class reported no cations detected, which was technically accurate but completely unhelpful. Always check your concentration before declaring a negative. The method also struggles with ions that form similar colored precipitates or have overlapping solubility products. Copper sulfide is black. Lead sulfide is black. Without careful sequential separation and confirmatory testing, you could easily misidentify one for the other. This is why the group scheme exists — it's not arbitrary, it's built around these exact overlaps. For mixtures with more than four or five ions at comparable concentrations, classical qualitative analysis becomes unwieldy. Instrumental methods like atomic absorption spectroscopy or ion chromatography handle those cases in a fraction of the time with far less ambiguity. The wet chemistry approach is still valuable for teaching the underlying principles and for situations where instrumentation isn't available, but don't pretend it's the most efficient tool for complex real-world samples.
One practical note on record keeping: write down observations in real time, not after the fact. The color of a precipitate, the shade of a flame, the exact volume of reagent added — these details fade from memory quickly and revisionist note-taking introduces errors that are nearly impossible to catch later. A dirty notebook in qualitative analysis is worse than no notebook at all. If you're setting up a lab session, make sure each student or pair has at least nine or twelve standard test tubes, a rack, small beakers for waste, pipettes or droppers that won't cross-contaminate, and access to a fume hood for anything involving H2S or concentrated acids. The minimum viable setup works, but cramped conditions with shared reagent bottles across multiple groups is a reliable way to generate contaminated samples and confused results.

Reading Results Like Someone Who Has Done This Before
A white precipitate with HCl means Group 1 is present, but it doesn't tell you which member. Silver chloride is white and curdy. Lead chloride is also white but more crystalline, and crucially it redissolves in hot water while silver chloride does not. That solubility difference is the actual confirmatory test, not just another reagent addition. For the confirmatory test with chromate, adding potassium dichromate to a Group 1 filtrate should give you a yellow precipitate if lead is present. But if your solution still has significant acidity from the initial HCl precipitation step, the chromate can convert to dichromate and the precipitate won't form cleanly. Neutralize or nearly neutralize before doing the chromate test. It's a small detail that separates a clean positive from a suspicious cloudy solution that leaves you guessing. Gas evolution tests need to be done right. Hydrogen sulfide from sulfide salts reacts with dilute acid to produce H2S gas, which turns lead acetate paper black. But carbonates also evolve gas with acid — CO2 — and if you're not careful about which gas you're actually testing for, you'll confuse a carbonate interference with a sulfide result. The lead acetate test is specific to H2S, but you should confirm by the odor description (rotten egg, though honestly you shouldn't be smelling it directly) and by the fact that CO2 won't blacken lead acetate paper. In a teaching lab, using thioacetamide instead of direct H2S is safer and gives the same sulfide precipitation under controlled hydrolysis conditions.
When everything fails, go back to fundamentals. Is your reagent actually fresh? Is your unknown actually what you think it is? Have you followed the sequence without skipping steps? Three of these four questions resolve the majority of problems that look like experimental failure. The other quarter is usually just a wrong concentration or a contaminated sample from the start. I'll leave it at that. There's a lot more to unpack on individual ion tests and tricky mixed-anion samples, but this covers the parts that tend to trip people up most consistently.