Validating Processes Without Losing Your Mind
Process validation is one of those topics that gets simplified into infographics and lost in translation. The core requirement hasn't changed much since the FDA issued their guidance back in 2011 — it's still about collecting evidence that a process consistently produces its intended result. The real question is how you actually build that evidence without turning your entire facility into a paper factory. I spent most of the early part of my career doing this work manually, which means I watched the industry shift from what they called "three-batch validation" to the lifecycle approach outlined in the Quality Management Systems Process Validation Guidance. There's a reason that shift happened. Three-batch validation worked fine for straightforward sterilization cycles or tablet compression. It fell apart fast when you had aseptic processing with media fills that required 3000 units per run and a six-month observation period before you could even assess compendial sterility.
Understanding the Quality Management Systems Process Validation Guidance Framework
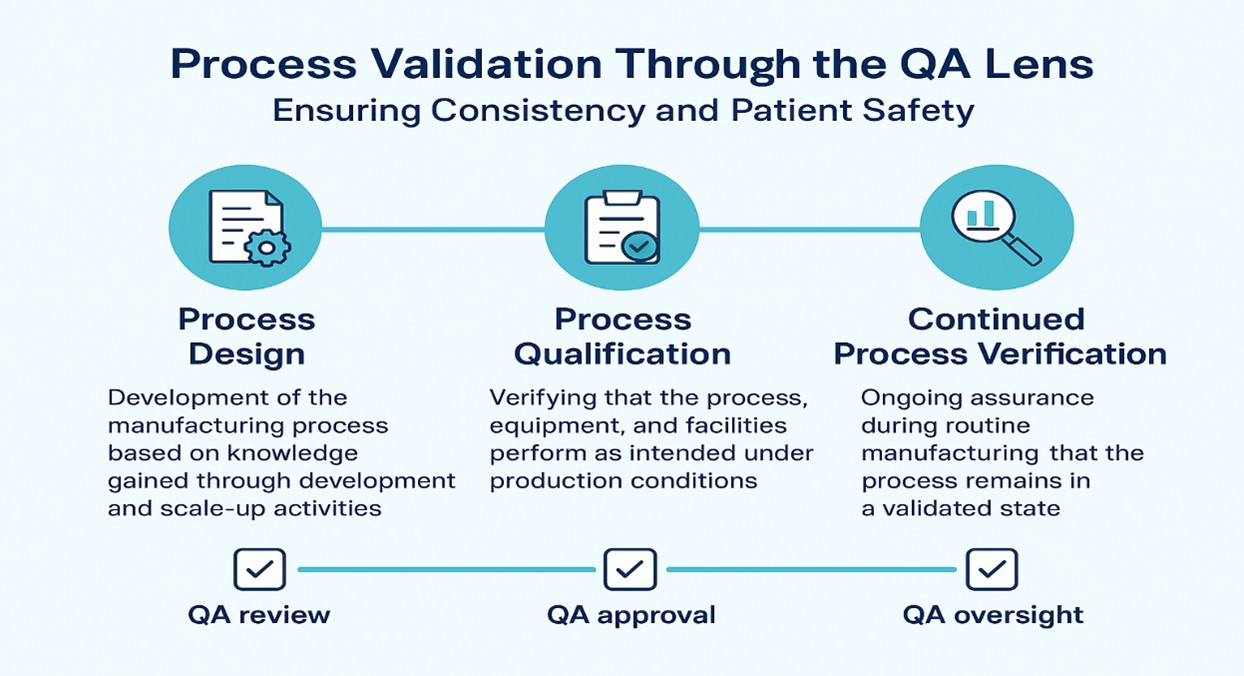
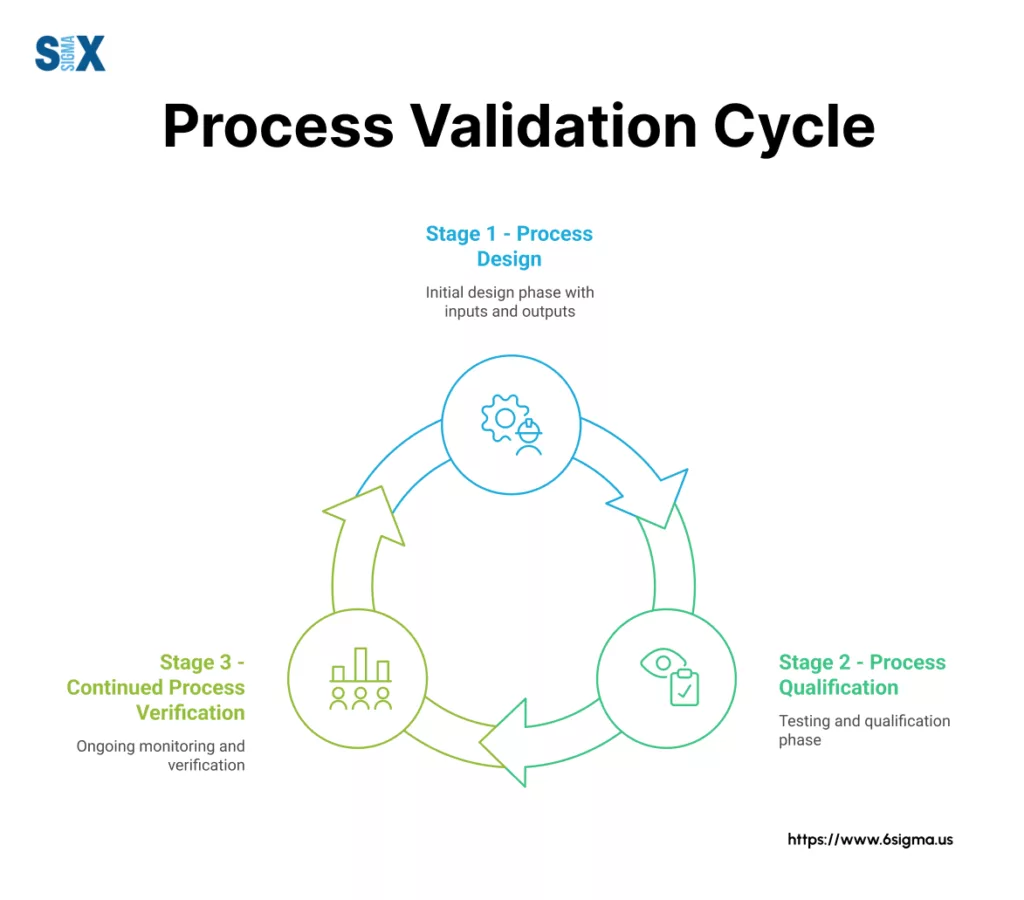
The lifecycle framework has three stages. Stage one is process design, where you define the process based on knowledge gained from development and scale-up activities. Stage two is process qualification, which confirms that the design is achieving the intended results during routine operation. Stage three is continued process verification, where you maintain control during commercial production through ongoing monitoring and trend analysis. Most people treat these stages as checkboxes. They aren't. The guidance is explicit that knowledge gained at each stage feeds into the next, and that you should adjust your approach based on risk. A simple unit operation like capsule filling doesn't need the same depth of characterization as a recombinant protein purification train. The guidance says this. The reality is that auditors sometimes ignore the nuance and expect identical documentation rigor across everything.
What Actually Gets Written Down
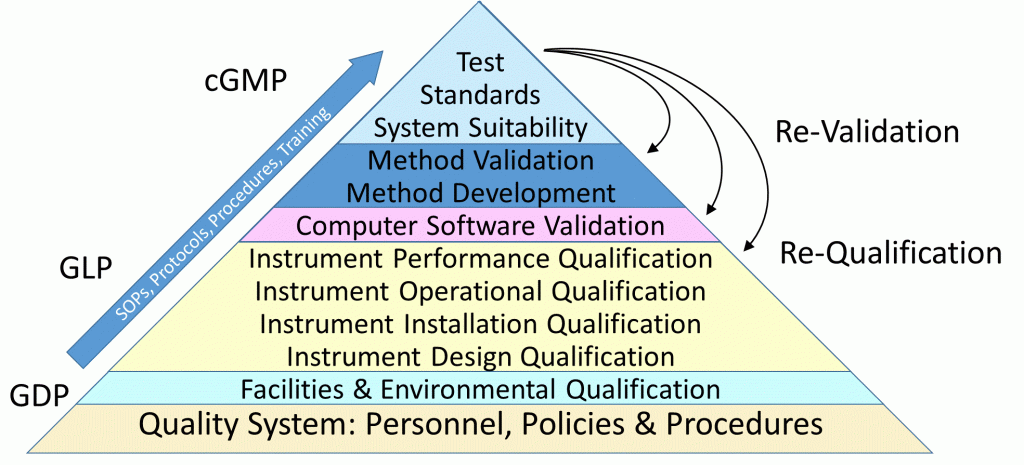
Your process design stage should produce a protocol that documents the process flow, critical quality attributes, critical process parameters, and the rationale for how each parameter was selected. This isn't a form letter. I've seen companies generate 200-page process design reports where half of it was copied from earlier projects with the product name changed. Auditors can spot this. More importantly, it defeats the purpose because you haven't actually characterized your specific process. The process qualification stage has two parts: installation qualification, operational qualification, and performance qualification. Installation qualification verifies that the equipment was installed correctly according to specifications. Operational qualification demonstrates that the equipment operates within defined limits. Performance qualification proves that the process works consistently under normal operating conditions using the intended materials and procedures. Here's the part most people skip. Performance qualification isn't just running three batches and calling it done. You need to demonstrate that your process is capable, which means showing statistical control. If your critical process parameter has a Cp of 0.8, you're not validated. You're just lucky so far.
Get the Full Details

Real Problems You'll Hit
I ran into a situation once where a fill volume process passed all three qualification batches within specification, but the Continued Process Verification data over the next six months showed a clear upward drift. The process was technically in control during qualification, but the control limits were set so wide that natural variation looked acceptable during the short PQ window while still producing borderline units during steady state. We caught it because the second shift technician flagged it in the daily log before anyone else reviewed the statistical output. The workaround wasn't dramatic. We tightened the process parameters during routine operation based on the CPV trending data, which brought the process back into a tighter band. We also recalibrated the fill machine calibration intervals from quarterly to monthly because the original schedule had been based on generic manufacturer recommendations rather than our observed drift pattern. This is the kind of thing the guidance anticipates in Stage 3, but it requires you to actually read the data instead of filing it away.
What the Guidance Gets Wrong or Leaves Ambiguous
The 2011 guidance was a significant improvement over the older framework, but it has gaps. It doesn't adequately address continuous manufacturing, which is increasingly common in the industry. It also doesn't provide clear criteria for when you need to revalidate versus when normal change control is sufficient. The line between a major change and a minor change can be subjective, and the guidance defers that to your scientific judgment, which is both correct and frustrating when an auditor disagrees with your assessment. Another blind spot is the treatment of software validation within the process validation lifecycle. Modern processes run on complex control systems with nested algorithms. The guidance mentions computerized systems but doesn't deeply integrate software validation requirements with process validation requirements. I've seen this create duplication where the same parameter gets validated twice under two different frameworks with slightly different criteria, which confuses operators and creates conflicting records.
Practical Approach That Works
Start with a risk assessment. Not a formal FMEA with a team of ten people and a two-day workshop. A focused, practical risk assessment that identifies your critical material attributes, critical process parameters, and critical quality attributes. Use tools you already have — process maps, historical data, operator interviews. Don't build a new methodology for this. Define your acceptance criteria before you run any batches. I know this sounds obvious, but I've seen qualification protocols where the acceptance criteria were defined after the data was collected, sometimes adjusted after seeing that the first run was borderline. That's not validation. That's post-hoc justification. Invest in your CPV program. This is where most companies fail. They treat Stage 3 as administrative overhead rather than a live control system. Set up your control charts with Western Electric rules or equivalent. Establish trending protocols that are reviewed on a scheduled basis, not just when an audit is coming. Flag and investigate any outliers with the same rigor you'd apply to a deviation.
Document your process knowledge. The guidance emphasizes that process understanding is the foundation. Keep records of your design space decisions, your scale-up rationale, your failure modes. When a regulatory inspection happens, they won't ask whether your forms are signed and dated. They'll ask why you set that parameter at that value and whether you have evidence to support it.
When Process Validation Doesn't Apply the Same Way
Not every process fits the traditional batch validation model. Biological products with living cells have inherent variability that makes statistical control challenging. Some processes are inherently heterogeneous. In these cases, you may need alternative approaches like patient-specific testing or real-time release testing, which shift the validation burden from the manufacturing process to in-process measurement and controls. These approaches require more investment in analytical method validation and process monitoring technology. They also require a different regulatory conversation. If you're considering moving away from traditional batch validation, talk to your regulatory contact early rather than building a complete alternative framework and presenting it as a finished product during inspection. The Quality Management Systems Process Validation Guidance is a living document in practice, even if the written guidance itself is fixed. Your validation strategy should evolve with your process knowledge. The companies that treat it as a one-time compliance exercise are the ones getting warning letters. The ones that use it as a framework for continuous improvement tend to run cleaner facilities and have easier inspections. Neither outcome is guaranteed, but the difference is real.