Titration for vinegar acidity — the real workflow
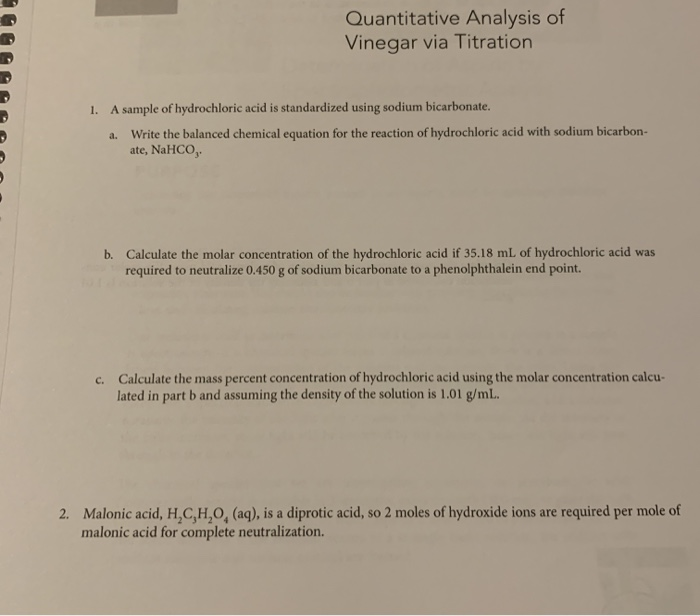
You take a measured volume of vinegar, dilute it in a flask, add a couple drops of phenolphthalein, and titrate against standardized sodium hydroxide until the faint pink persists for thirty seconds. That is the Quantitative Analysis Of Vinegar Via Titration in its simplest form. Most people stop there. They get the calculation right on paper and then proceed to ruin their results in the lab. The acid in vinegar is acetic acid, CH3COOH, a monoprotic weak acid. The reaction with sodium hydro CO3 + NaOH CH3COONa + H2O goes to completion, and the stoichiometry is straightforward. One mole of NaOH neutralizes one mole of acetic acid. The math is not the hard part. Getting a reliable endpoint is.
Standardizing your NaOH before you touch the vinegar
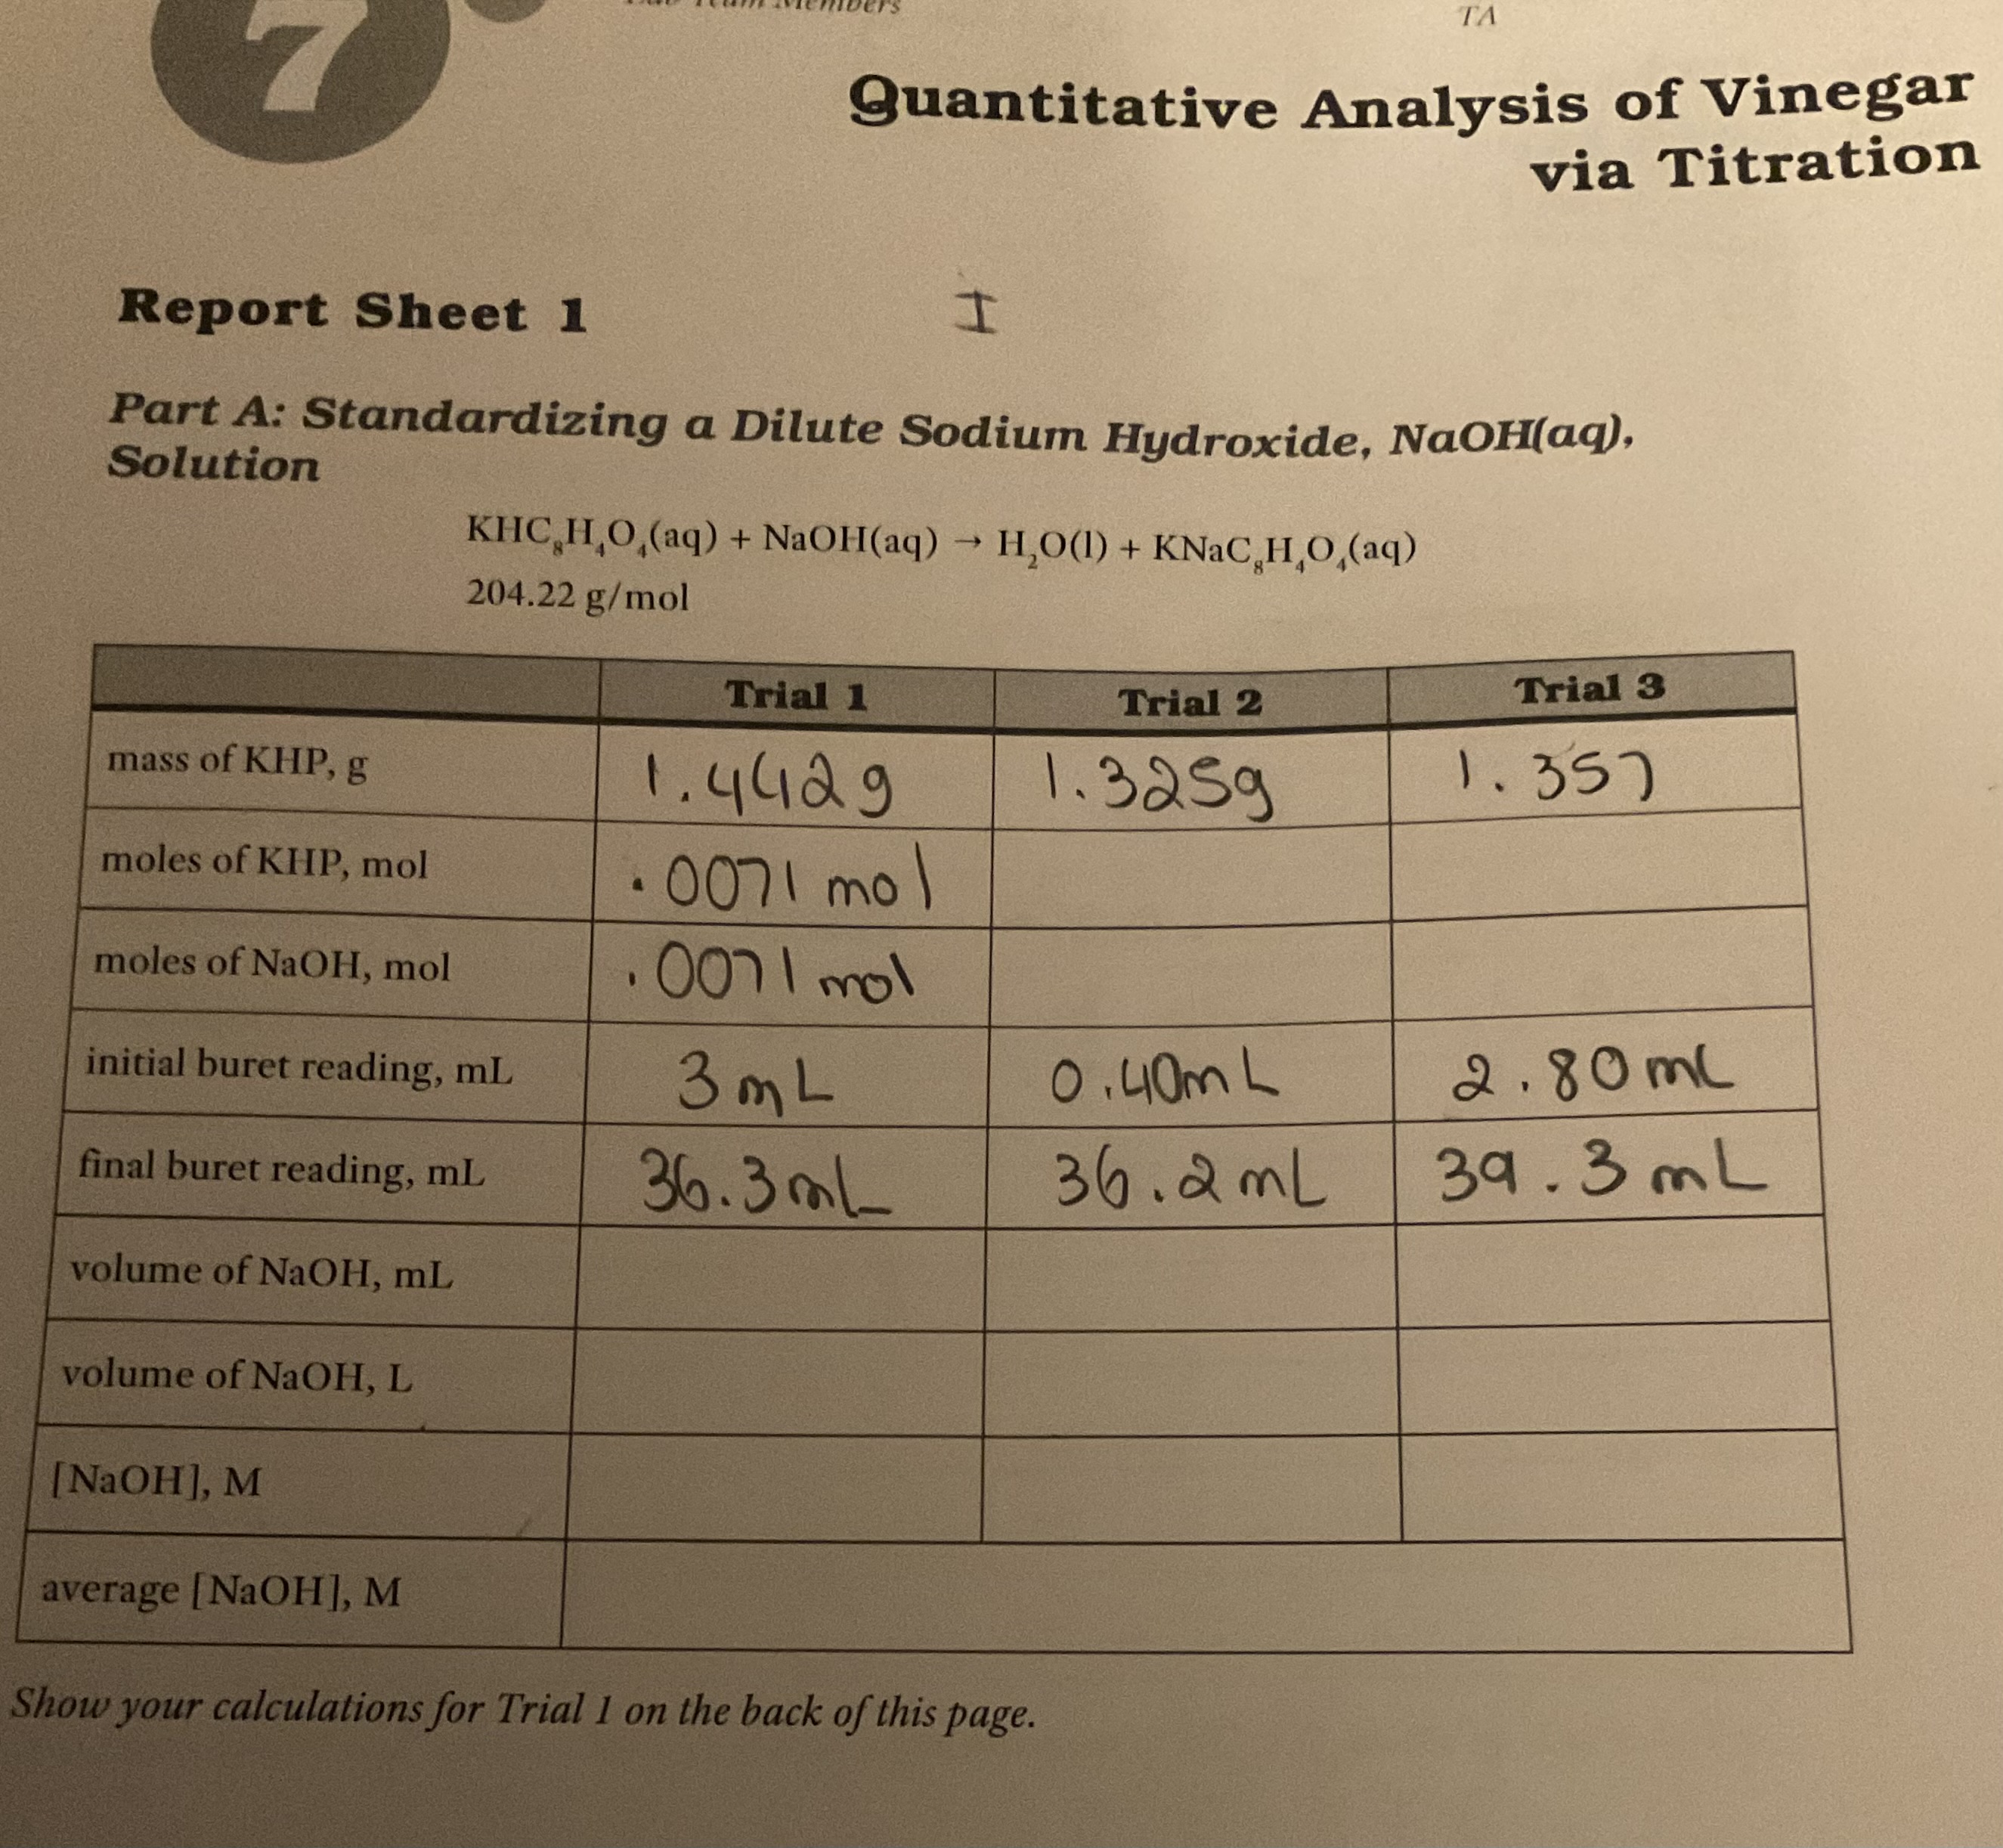
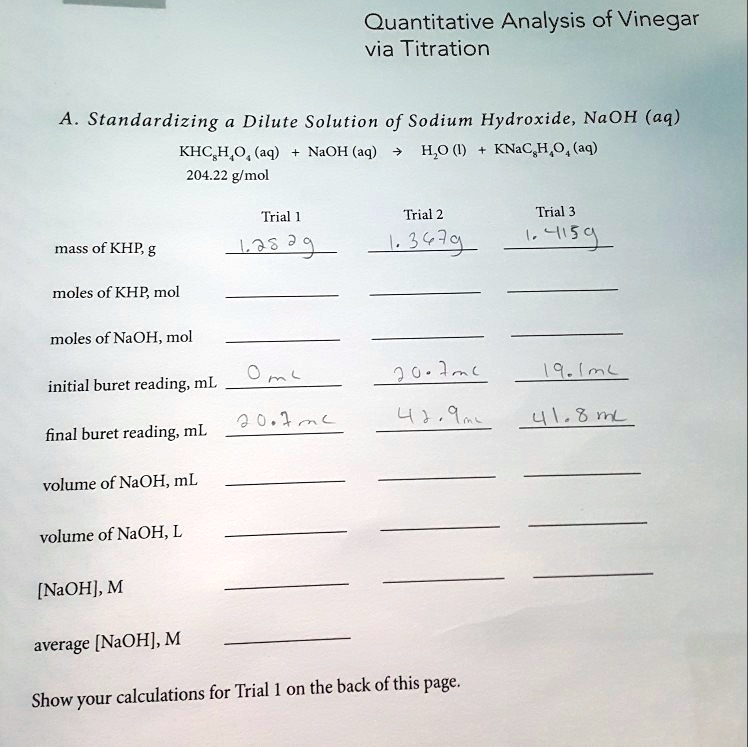
This is where people go wrong, and it is the single biggest source of error in undergraduate labs and small-scale quality control work. Sodium hydroxide absorbs moisture from the air and carbon dioxide as well. A bottle labeled 0.1 M NaOH is not 0.1 M NaOH after it has sat open for a week. You must standardize against a primary standard. Potassium hydrogen phthalate, KHP, is the usual choice. It is stable, available as a high-purity solid, and has a molecular weight of 204.22 g/mol. Here is how I do it. Dry your KHP at 110 degrees Celsius for two hours, cool it in a desiccator, and weigh out approximately 0.4 to 0.5 grams into an Erlenmeyer flask. Dissolve in about fifty milliliters of CO2-free distilled water. I make CO2-free water by boiling distilled water for five minutes and letting it cool under a layer of mineral oil or in a sealed container. Titrate with your NaOH solution using phenolphthalein. The endpoint is pale pink, not hot magenta. Calculate the exact molarity from the mass of KHP and the volume of NaOH used. Do this on the same day you run your vinegar samples.
What happens at the endpoint and why it matters
Phenolphthalein changes color in the pH range of about 8.2 to 10. The equivalence point for acetic acid titrated with NaOH sits around pH 8.7 to 9.2, depending on concentration. This means phenolphthalein is actually a reasonable indicator for this reaction. Methyl orange would be wrong because it changes color around pH 3.5 to 4.5, well before the equivalence point. Using the wrong indicator can throw your result off by several percent. But here is something most guides do not mention. The endpoint you see with phenolphthalein is slightly past the true equivalence point. The acetate ion that forms is a weak base, and it hydrolyzes in water to produce a small amount of hydroxide. This is normal. It is built into the chemistry. What is not normal is drifting your endpoint by chasing a deeper pink color. If you are adding half a drop of NaOH and the solution flips from colorless to dark pink, you have overshot. The result will be too high. Practice making the last few milliliters of titrant in quarter-drop increments. Rinse the sides of the flask with deionized water from a wash bottle during the titration so that any reagent clinging to the glass gets incorporated into the reaction volume.
Get the Full Details
Dealing with colored vinegar samples
I once ran Quantitative Analysis Of Vinegar Via Titration on a batch of aged balsamic vinegar that was essentially opaque brown. Phenolphthalein was invisible. The pink endpoint could not be detected visually at all. I switched to a pH meter and did a potentiometric titration instead. I placed a combined glass electrode in the flask, set up a burette with 0.1 M NaOH, and recorded the pH after every 0.5 mL addition. The inflection point on the pH versus volume curve gave me the equivalence volume clearly. The Gran plot method works well here if you need higher precision. I found the first derivative of the titration curve and took the peak. This took about twenty minutes per sample instead of five, but it was the only way to get a readable result. If you are working with regular white vinegar, this problem does not exist. The sample is clear and the phenolphthalein endpoint is easy to see. But do not assume every commercial vinegar is clear. Flavored vinegars, wine vinegars, and particularly artisanal or aged products can be deeply colored. A quick visual check before you start titrating saves you from wasting a full titration cycle and having to figure out why your volume readings make no sense.
Calculations and common pitfalls
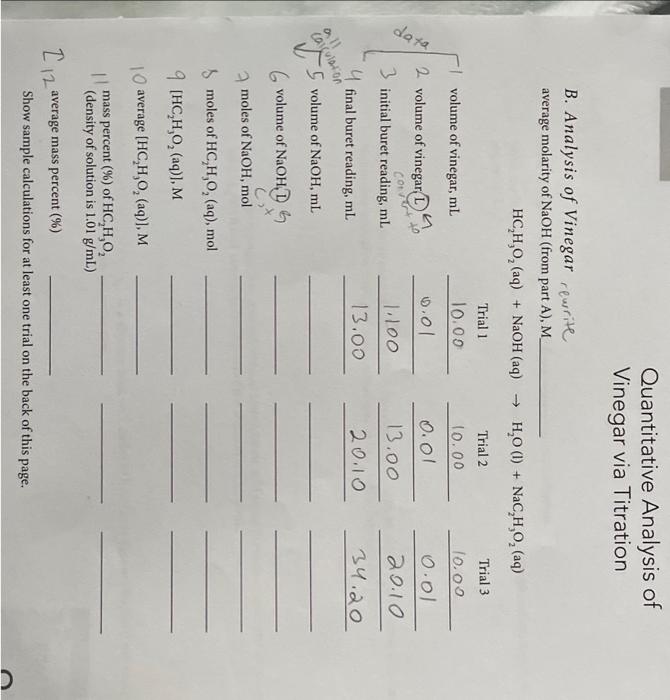
The core equation is simple. Moles of NaOH equals molarity times volume in liters. Moles of acetic acid equals moles of NaOH because of the 1:1 stoichiometry. Mass of acetic acid equals moles times the molecular weight of acetic acid, which is 60.05 g/mol. Percent mass of acetic acid equals the mass of acetic acid divided by the mass of the vinegar sample, multiplied by 100. A typical white vinegar sample is about 5 percent acetic acid by mass, which translates to roughly 0.83 M. One mistake that comes up constantly is using the volume of the diluted sample instead of the original undiluted aliquot in the final calculation. If you pipette 10 mL of vinegar and dilute it to 50 mL with water before titrating, your calculation must use the original 10 mL, not the 50 mL. The water is just a solvent. It does not change the number of moles of acetic acid present. Forgetting this multiplies your error by five. Another issue is temperature. Solution volumes expand and contract with temperature. A burette calibrated at 20 degrees Celsius will read slightly off at 25 degrees. The effect is small but measurable. A 5 mL volume change from a 5-degree temperature shift is about 0.025 mL, which introduces roughly a 0.05 percent error in a typical titration. It is negligible for most teaching labs but noticeable if you are doing high-precision work or quality control for food manufacturing. More importantly, the dissociation constant of acetic acid changes with temperature, which shifts the equivalence point pH slightly. Again, this is a minor effect for routine analysis but worth knowing if your lab environment is uncontrolled.
When titration is not the right tool
Titration works well for acetic acid concentrations above about 0.01 M. Below that, the endpoint becomes vague and indicator errors dominate. Vinegar samples with very low acidity, such as certain cleaning solutions labeled "diluted vinegar," may fall below this range if you do not adjust your titrant concentration or sample volume. If the acetic acid content is below 0.5 percent, consider using a more dilute NaOH solution, perhaps 0.01 M, and increasing your sample aliquot to 25 or 50 mL. You will use more titrant and the titration will take longer, but the endpoint will be sharper relative to the volume delivered. There are also samples where titration gives misleading results. Vinegars containing significant amounts of other acids, such as citric acid added as a preservative or phosphoric acid in some industrial cleaning vinegars, will report total titratable acidity rather than acetic acid specifically. The result will be accurate for total acidity but inflated if you assume all the acid is acetic acid. Gas chromatography or enzymatic assays are better choices when you need speciated acid analysis. For routine quality checks on standard table vinegar, titration remains fast, cheap, and sufficiently accurate.
Practical tips from actual lab work
Use class A volumetric glassware if precision matters. A class A 25 mL pipette has a tolerance of ±0.03 mL. A class B pipette allows ±0.06 mL. The difference is small but accumulates across multiple measurements. A 50 mL burette graded in 0.1 mL increments lets you estimate to ±0.02 mL with practice. Reading the meniscus at eye level is mandatory. Parallax error alone can add 0.05 mL of uncertainty, which is a full percent in a typical titration. Rinse your burette with the titrant solution before filling it, not just with water. Residual water in the burette dilutes the NaOH and changes its effective concentration mid-titration. Do the same with your pipette. Rinse it with the sample solution before drawing up the vinegar. These steps take ten seconds and prevent a class of systematic error that is harder to detect than random measurement noise. Record your initial and final burette readings to two decimal places. Even if your burette is graduated in 0.1 mL increments, you should estimate the position between marks. A reading of 23.47 mL is different from 23.50 mL, and in a tight titration where the volume difference between trials might be 0.1 mL, that estimation matters. Repeat each titration at least twice, ideally three times, and report the average. If your duplicate results differ by more than 0.1 mL, redo the titration. Something went wrong, and you do not want to build a report on bad data.
Common equipment and reagent sources
Standard laboratory glassware suppliers carry burettes, pipettes, and Erlenmeyer flasks suitable for this work. Titration stands with magnetic stirrers simplify the process significantly compared to manual swirling. A magnetic stir bar spinning at moderate speed gives consistent mixing without the variability of hand swirling, and it reduces the chance of overshooting the endpoint. For NaOH, buy solid pellets and prepare your own stock solution rather than relying on pre-diluted bottles. For KHP, Thermo Fisher, Sigma-Aldrich, and VWR all sell primary standard grade material certified for titration work. Phenolphthalein indicator solution is widely available from any chemistry supply company. Store it in a dark bottle because light degrades it over time. This method will give you acetic acid content within about one to two percent relative error if you follow the procedure carefully. That is adequate for most educational purposes and routine commercial quality checks. If you need better precision, you will need to control temperature more rigorously, use automated potentiometric titration, and account for activity coefficients rather than treating the solution as ideal. Most vinegar analysis does not require that level of rigor. Knowing when the simple version is sufficient and when it is not is the actual skill here.