Quantum Mechanical Model Definition Chemistry
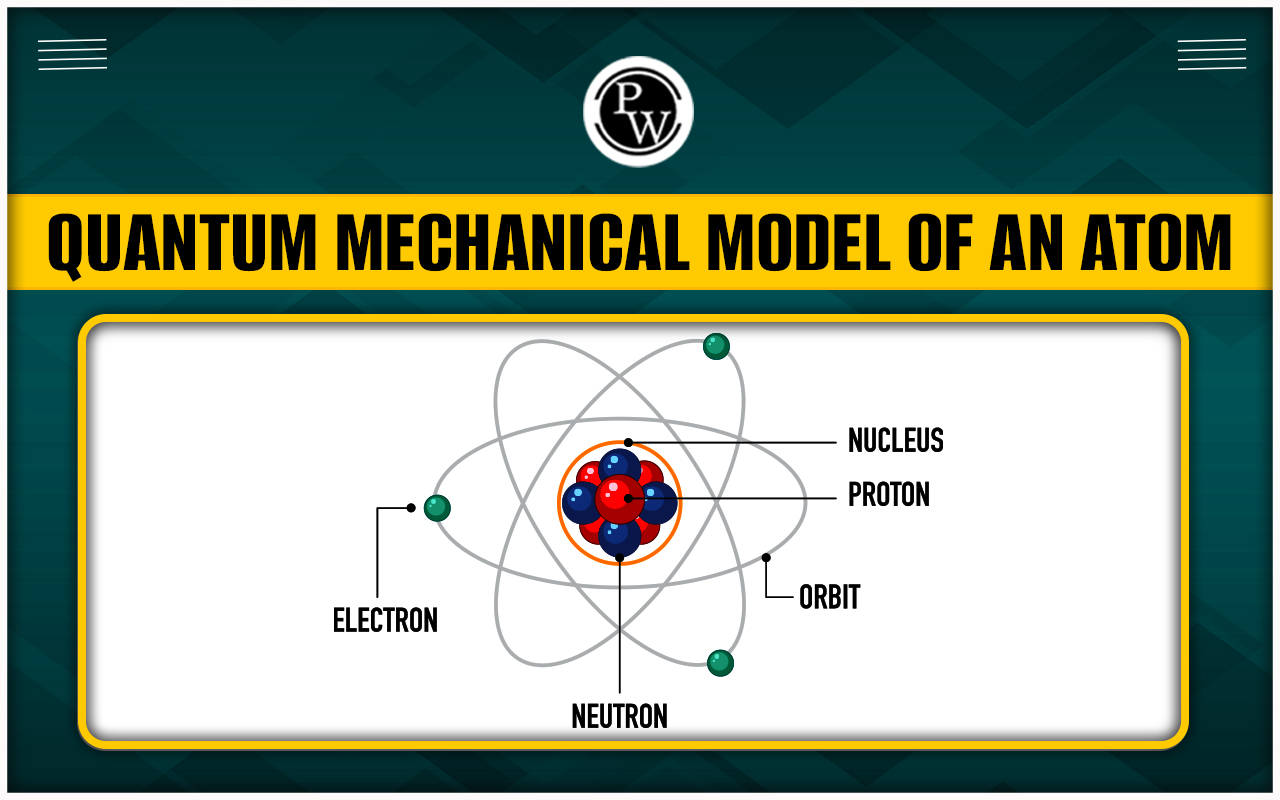
The quantum mechanical model of the atom is the framework we actually use to predict how electrons behave in atoms. It replaced the Bohr model because the Bohr model only worked for hydrogen and failed the moment you tried to apply it to anything with more than one electron. That was a real problem in the lab. You could not fit experimental data to a planetary orbit picture, and students kept trying to draw electrons circling nuclei like tiny planets. It does not work that way.
What the Quantum Mechanical Model Definition Chemistry Actually Means
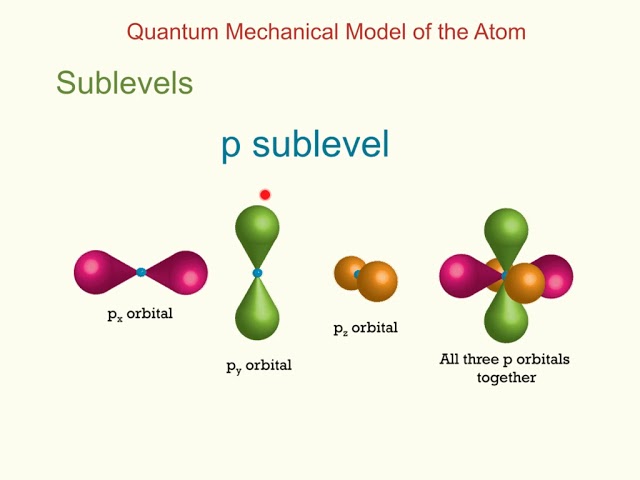
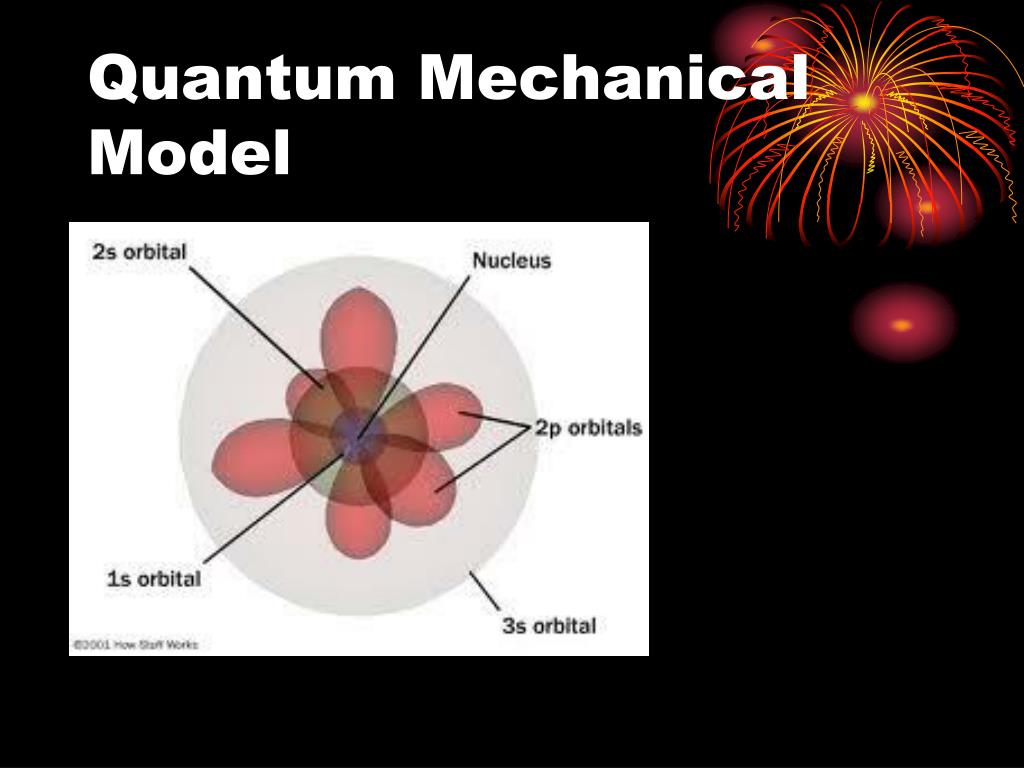
In practice, the quantum mechanical model treats electrons as wavefunctions rather than particles traveling on fixed paths. The Schrödinger equation is the engine behind it. You input a nuclear potential and an electron, and the math returns a wavefunction. From that wavefunction you get probabilities, not trajectories. The model gives you orbitals instead of orbits. An orbital is just a region of space where the probability of finding an electron is above a certain threshold, usually 90 percent.I have seen people confuse the three-dimensional shapes of p orbitals with literal solid objects. They are probability clouds. Saying an electron is in a dumbbell-shaped region is shorthand for saying the integrated probability density over that region comes out to roughly 90 percent. It is not a physical boundary. It is a contour map of likelihood. That distinction matters when you start doing actual calculations or interpreting spectroscopic data.
How the Model Actually Works in Practice
The model breaks down into quantum numbers, and you need to track all four. The principal quantum number determines the shell and roughly the energy level. The angular momentum quantum number determines the subshell shape: s, p, d, or f. The magnetic quantum number sets the orientation in space. The spin quantum number is either plus or minus one half. Every electron in an atom gets a unique set of those four numbers because of the Pauli exclusion principle. That is how electron configurations come out the way they do.When you are working with multi-electron atoms, the energy ordering is not strictly by principal quantum number. You have to account for shielding and penetration effects. An s orbital penetrates closer to the nucleus than a p orbital in the same shell, so it sits lower in energy. That is why the 4s orbital fills before the 3d orbital in potassium and calcium. Once you start filling d orbitals, the energy levels shift again. The Aufbau principle works as a rough guide, but you will encounter exceptions. Chromium and copper are the classic ones, and there are more in the later transition metals and lanthanides.
A Real Edge Case That Nearly Broke My Workflow
A few years back I was assigning electron configurations for a series of late transition metal complexes, and the standard n plus l rule predicted configurations that did not match the observed magnetic data. I was modeling a cobalt complex where the ligand field splitting was small enough that high-spin and low-spin states were close in energy. The quantum mechanical model told me the expected configuration, but the actual ground state depended on subtle exchange energy and spin-orbit coupling effects that the basic model does not capture directly. I had to switch from a simple configuration lookup to running a DFT calculation with an appropriate functional. That took about twenty minutes per geometry optimization on a modern workstation, versus the few seconds a textbook prediction would have given me. I learned to flag any third-row transition metal or lanthanide complex where magnetism matters and run the computational check immediately rather than trusting the simplified rules.Get the Full Details

Where the Model Falls Short
The quantum mechanical model is powerful, but it is not a complete solution for everything. Solving the Schrödinger equation exactly is only possible for single-electron systems like hydrogen and hydrogen-like ions. For anything with more than one electron, you need approximations. Hartree-Fock, density functional theory, post-Hartree-Fock methods like MP2 or CCSD(T) are the standard tools, and each has trade-offs. Hartree-Fock ignores electron correlation entirely. DFT includes correlation approximately, but the choice of functional can change your results enough to flip a predicted reaction barrier. Post-Hartree-Fock methods get expensive fast. Coupled cluster scales steeply with system size, so a 50-atom organic molecule can take hours on a good cluster while a smaller transition metal complex might take days. If you are only looking at basic atomic structure for an introductory chemistry course, the quantum mechanical model definition chemistry is sufficient. You learn orbitals, quantum numbers, and how to build configurations. If you are actually using the model to predict properties, you need to know which approximation you are relying on and what errors it introduces. For quick molecular orbital sketches, Hückel theory works fine for conjugated hydrocarbons, but it breaks down the moment heteroatoms or three-dimensional geometry enter the picture. For those cases, you move to ab initio or DFT software. Gaussian, ORCA, and Q-Chem are the common choices. ORCA is free for academic use and handles a broad range of methods without requiring a commercial license.Common Pitfalls to Avoid
Students regularly mix up nodes with orbital boundaries. A node is just a region where the wavefunction passes through zero. It is not a physical wall. Another frequent mistake is assuming that orbital shape implies electron speed. Electrons do not orbit faster near the nucleus because they are whizzing around. The probability density changes because of the mathematical form of the wavefunction. The kinetic energy operator in the Hamiltonian captures the curvature of the wavefunction, and high curvature near the nucleus corresponds to higher kinetic energy contribution, not classical orbital velocity. You should also stop treating the quantum mechanical model as if it gives exact answers. Every result carries approximation error. When a paper reports a bond length within a few picometers of experiment, that is usually DFT with a decent functional and a reasonably large basis set. When a textbook shows orbital diagrams with perfectly filled and empty levels, that is a pedagogical simplification. Real systems have thermal broadening, vibronic coupling, and environmental effects that shift energy levels. If your goal is to interpret experimental spectra, you need to model those effects explicitly rather than relying on the bare orbital picture.The model itself remains the foundation of modern chemistry. Without it, there is no rational explanation for periodic trends, bonding, or spectroscopy. The practical value comes from understanding what the model provides, what it approximates, and when you need to go beyond the basic definition into computational methods. That shift usually happens around the time you stop drawing orbitals by hand and start feeding coordinates into software, expecting results that actually match what happens in the lab.
Quantum Mechanical Model Definition Chemistry in Everyday Use
For most students and practicing chemists who do not run simulations daily, the quantum mechanical model definition chemistry reduces to knowing how to assign quantum numbers, read electron configurations, predict basic orbital overlap for bonding, and recognize when a problem requires computational input. You do not need to solve differential equations to use the model effectively. You do need to know its limits. When you hit those limits, you switch to the appropriate approximation and accept that the output carries uncertainty. That is how the model is actually used, not as a perfect predictor, but as a structured way to reason about electronic behavior until you need more precision.