Writing Rate Expressions for Kinetics Problems
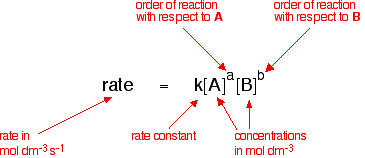
Most people encounter the rate of reaction equation in a second-year chemistry course and immediately treat it as a memorization task. It's not. You derive it from the rate law, and you choose the rate law based on experimental data, not intuition. Here's how it actually works in practice. The general form is rate = k[A]^m[B]^n where k is the rate constant, concentrations are molar, and the exponents come from experiment. That's the baseline. What nobody tells you is that the rate constant k carries different units depending on the overall order. Zero order: M/s. First order: 1/s. Second order: 1/(M·s). Third order: 1/(M²·s). If your units don't match, you've set up something wrong. I spent a good afternoon last year debugging a student lab report where the calculated k value was numerically correct but the units were completely inconsistent because they'd mixed mol/L with mmol/L halfway through. The final answer had the right digits but was off by a factor of a thousand in practice. The fix was just writing all concentrations in the same units before plugging anything in. Took thirty seconds once we found it.
Here's the practical sequence you should follow when you're actually working a problem: First, determine the reaction order. You do this by looking at how the initial rate changes when you vary one reactant's concentration while holding everything else constant. If doubling [A] doubles the rate, the reaction is first order in A. If doubling [A] quadruples the rate, it's second order. If nothing happens, it's zero order. This is the integral method approach and it's what you'll use in most course settings. Second, calculate k using the determined orders and any one set of experimental data. Don't average k values across trials unless your data is clean. If your k values vary by more than ten percent between trials, your rate law is probably wrong or there's contamination in the experiment. I've seen this happen when people reuse reaction vessels without proper rinsing and carry over catalytic impurities.
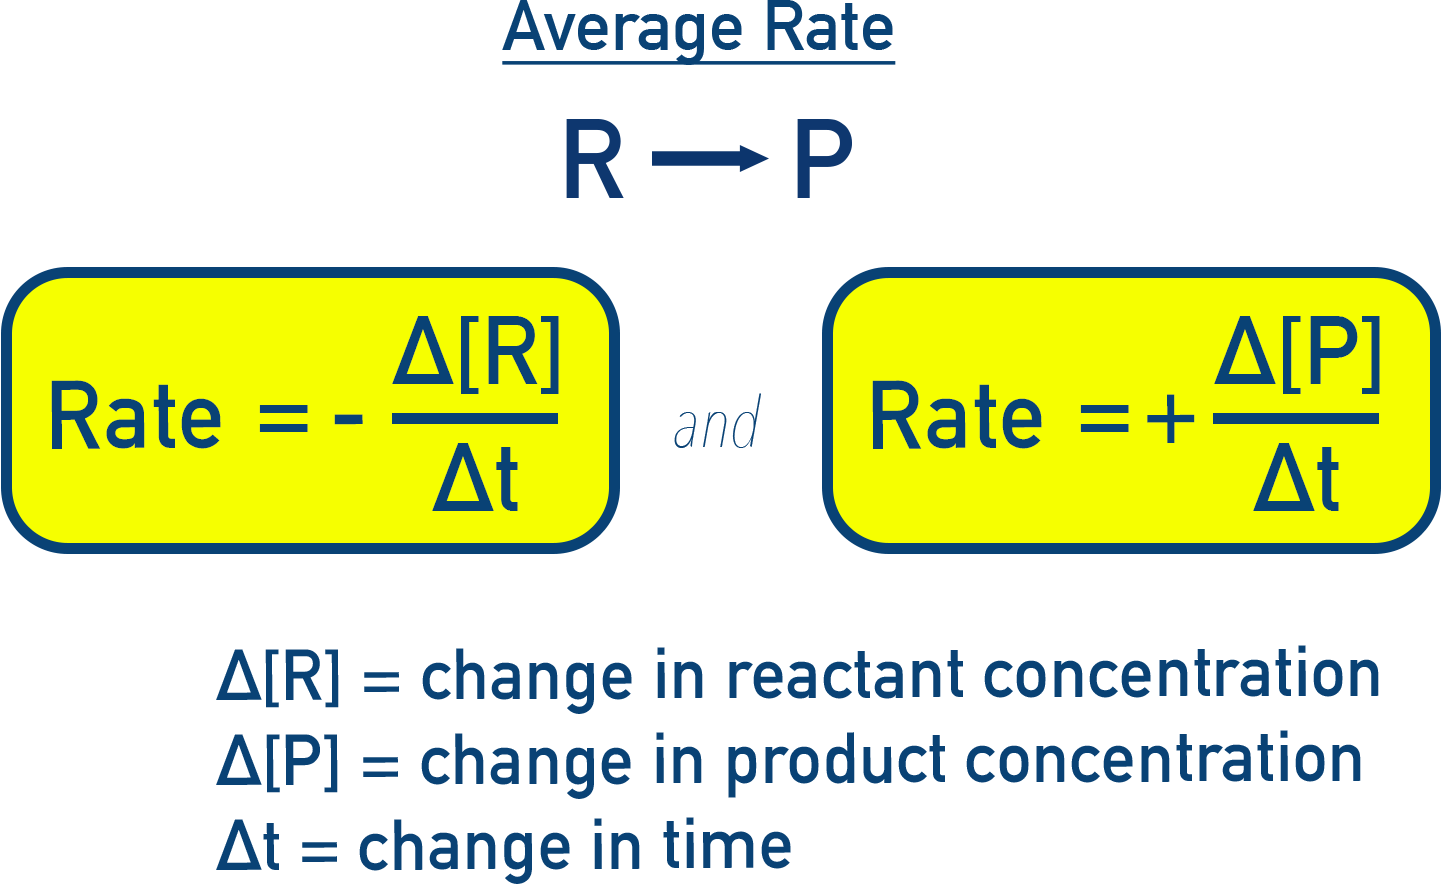
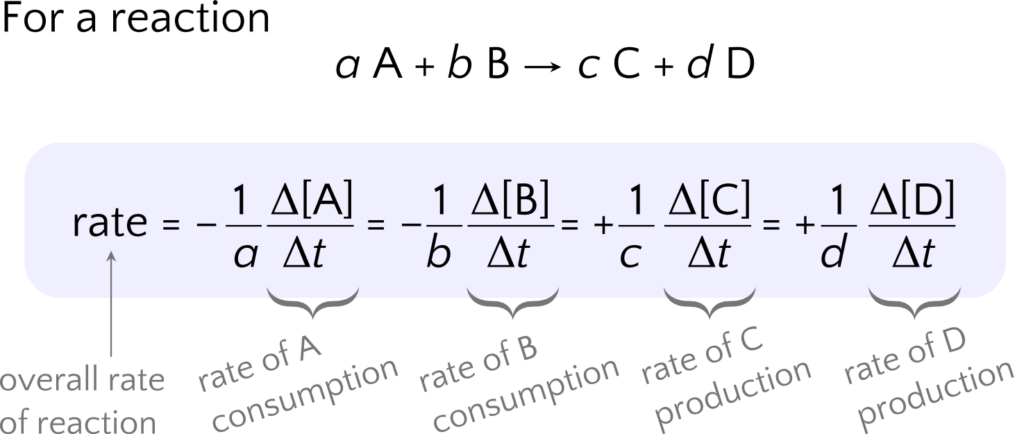
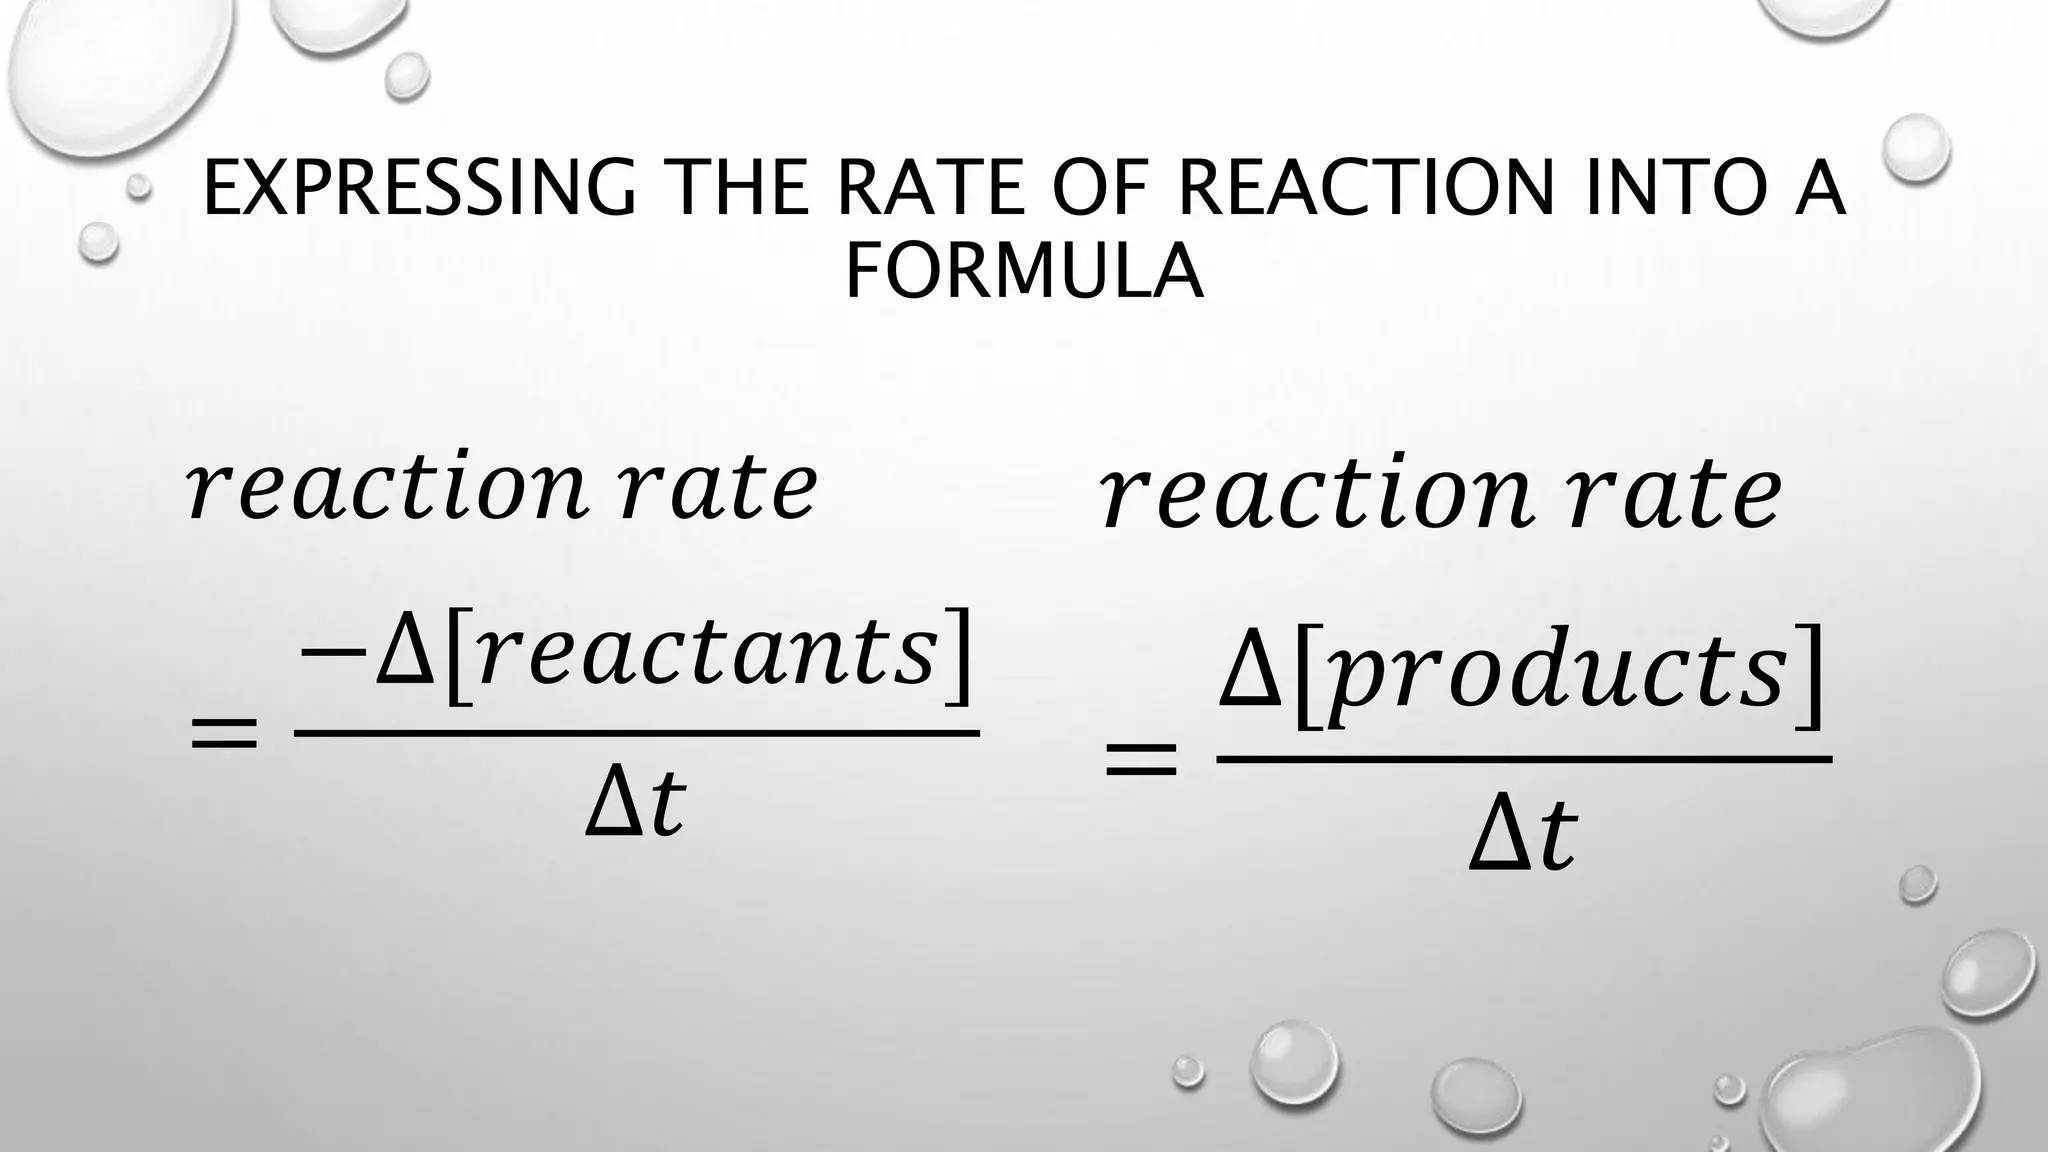
Third, write the full Rate Of Reaction Equation with your determined exponents and calculated k value, including units for k. That last step gets skipped constantly and it's the difference between a complete answer and one that's technically incomplete. There's a common misconception that the stoichiometric coefficients from the balanced equation directly become the exponents in the rate law. They don't, unless the reaction is an elementary step. For a multi-step mechanism, the exponents reflect the rate-determining step, not the overall stoichiometry. I've corrected this mistake dozens of times. A reaction like 2NO 4NO + O is first order overall, not second order, because the mechanism involves a unimolecular decomposition in its slow step. The coefficients are irrelevant to the rate law here. Another thing that catches people out: the rate of reaction equation can be written in terms of any reactant or product, but the numerical value depends on which species you track unless you normalize by stoichiometric coefficient. Rate = -d[A]/dt = -(1/2)d[B]/dt = d[C]/dt. Students often write expressions that look like equalities but actually differ by a factor of two or three depending on how they define "rate." Always specify which species' consumption or formation rate you're calculating, or use the normalized convention consistently.
Get the Full Details
The main limitation of this approach is that it only works cleanly for reactions with simple, well-defined kinetics. If your reaction has autocatalysis, enzyme saturation, or a complex mechanism with competing pathways, the simple rate = k[A]^m[B]^n form breaks down. In those cases you need integrated rate laws, the steady-state approximation, or numerical fitting. I usually recommend starting with the simple form anyway, checking residuals against your experimental data, and only moving to something more complex if the basic model consistently fails to fit within experimental error. For quick reference, here's a summary of the integrated forms you'll need depending on the order you determine: Zero order: [A] = [A] - kt. Plot [A] versus t, slope is -k.
First order: ln[A] = ln[A] - kt. Plot ln[A] versus t, slope is -k. Second order: 1/[A] = 1/[A] + kt. Plot 1/[A] versus t, slope is k. The quickest way to determine order in practice is to try all three plots with your data and see which one gives you a straight line. The R-squared value tells you immediately. I usually do this in a spreadsheet and it takes about five minutes for a standard dataset.
Temperature dependence is another layer you'll encounter. The rate constant follows the Arrhenius equation: k = Ae^(-Ea/RT). If you measure k at two or more temperatures, you can calculate the activation energy Ea. A typical rule of thumb is that reaction rates roughly double for every ten degree Celsius increase, but that's a rough heuristic. The actual factor depends on Ea. For a reaction with Ea = 50 kJ/mol, the rate increases by about 1.7x per 10°C near room temperature. For Ea = 100 kJ/mol, it increases by about 2.8x. The heuristic works for ballpark estimates but will mislead you if you need precision. If you're working with real experimental data and the simple models aren't fitting, the problem is usually one of three things: the reaction isn't elementary, you have side reactions or equilibrium effects contaminating your measurements, or your concentration range is too narrow to distinguish between similar rate laws. In my experience, the third issue is the most common. If all your concentrations fall within a ten percent window, first order and second order kinetics can look nearly identical. Spread your concentration range wider if possible, or use a statistical test like an F-test to compare model fits rather than eyeballing R-squared values. That's enough for getting started. The math is straightforward once you accept that the exponents come from data, not from balancing equations, and that the whole framework falls apart if your experimental design is sloppy.