Working With Amides in the Lab
Amides are annoying. They look like they should be reactive because they have a carbonyl, but that lone pair on nitrogen pushes electron density into the C=O and makes it way less electrophilic than a ketone or ester. You will spend a lot of time trying to get something to happen at room temperature and nothing happening, then finally boiling your reaction under reflux for six hours and getting 40% yield. This is normal. It is just what amides do. The core problem most people run into is that amides sit in a gray zone. They are stable enough to survive almost any workup, which is why they are used as protecting groups and in peptide synthesis. But that same stability means they refuse to participate in the reactions you want them to unless you push them hard. I learned this the hard way during a multi-step synthesis where I needed to convert a primary amide to a nitrile using SOCl2. The literature procedure said 2 hours at reflux in toluene. My reaction was still showing starting material by TLC after 8 hours. I switched to using POCl3 with a catalytic amount of DMF instead, which activated the amide oxygen through an iminium intermediate, and the conversion went to completion in under an hour at room temperature. The DMF trick changes the mechanism from a direct chlorination to something faster, and it saved me from having to heat the whole flask setup overnight.
Common Reaction With Amide Issues
Reduction of amides to amines with LiAlH4 is one of the most common transformations, but it is also where people make mistakes. Unlike ester reduction, amide reduction doesn't stop at an aldehyde. The mechanism goes through an iminium ion intermediate that is immediately reduced further. If you need that aldehyde, you have to use DIBAL-H at low temperature and quench carefully, usually at -78°C in toluene or CH2Cl2. Getting the temperature right matters more than anything else here. If your dry ice-acetone bath isn't actually at -78°C when you add the DIBAL-H, you will get a mixture of amine and aldehyde and a difficult purification. Acid-catalyzed hydrolysis of amides requires strong acid and heat. Concentrated HCl or H2SO4 at reflux for several hours is standard. The reaction is slow because the carbonyl carbon is not very electrophilic. You can speed it up by using a phase transfer catalyst or microwave irradiation if your lab has that equipment. Microwave-assisted hydrolysis in sealed tubes with 6M HCl at 180°C can complete in 15 to 30 minutes what normally takes 6 to 12 hours. I use this method when I need to check whether a hydrolysis worked before committing to a full-scale reaction. It cuts the waiting time dramatically. Basic hydrolysis works too but is generally slower and requires more forcing conditions. NaOH in ethanol-water at reflux is the typical setup. The product is a carboxylate salt and ammonia or an amine. You need to acidify the reaction mixture after completion to precipitate or extract the carboxylic acid. This is straightforward in principle but the emulsion problems during extraction can be real, especially if your substrate has any hydrophobic character. Adding brine and filtering through celite usually sorts it out.
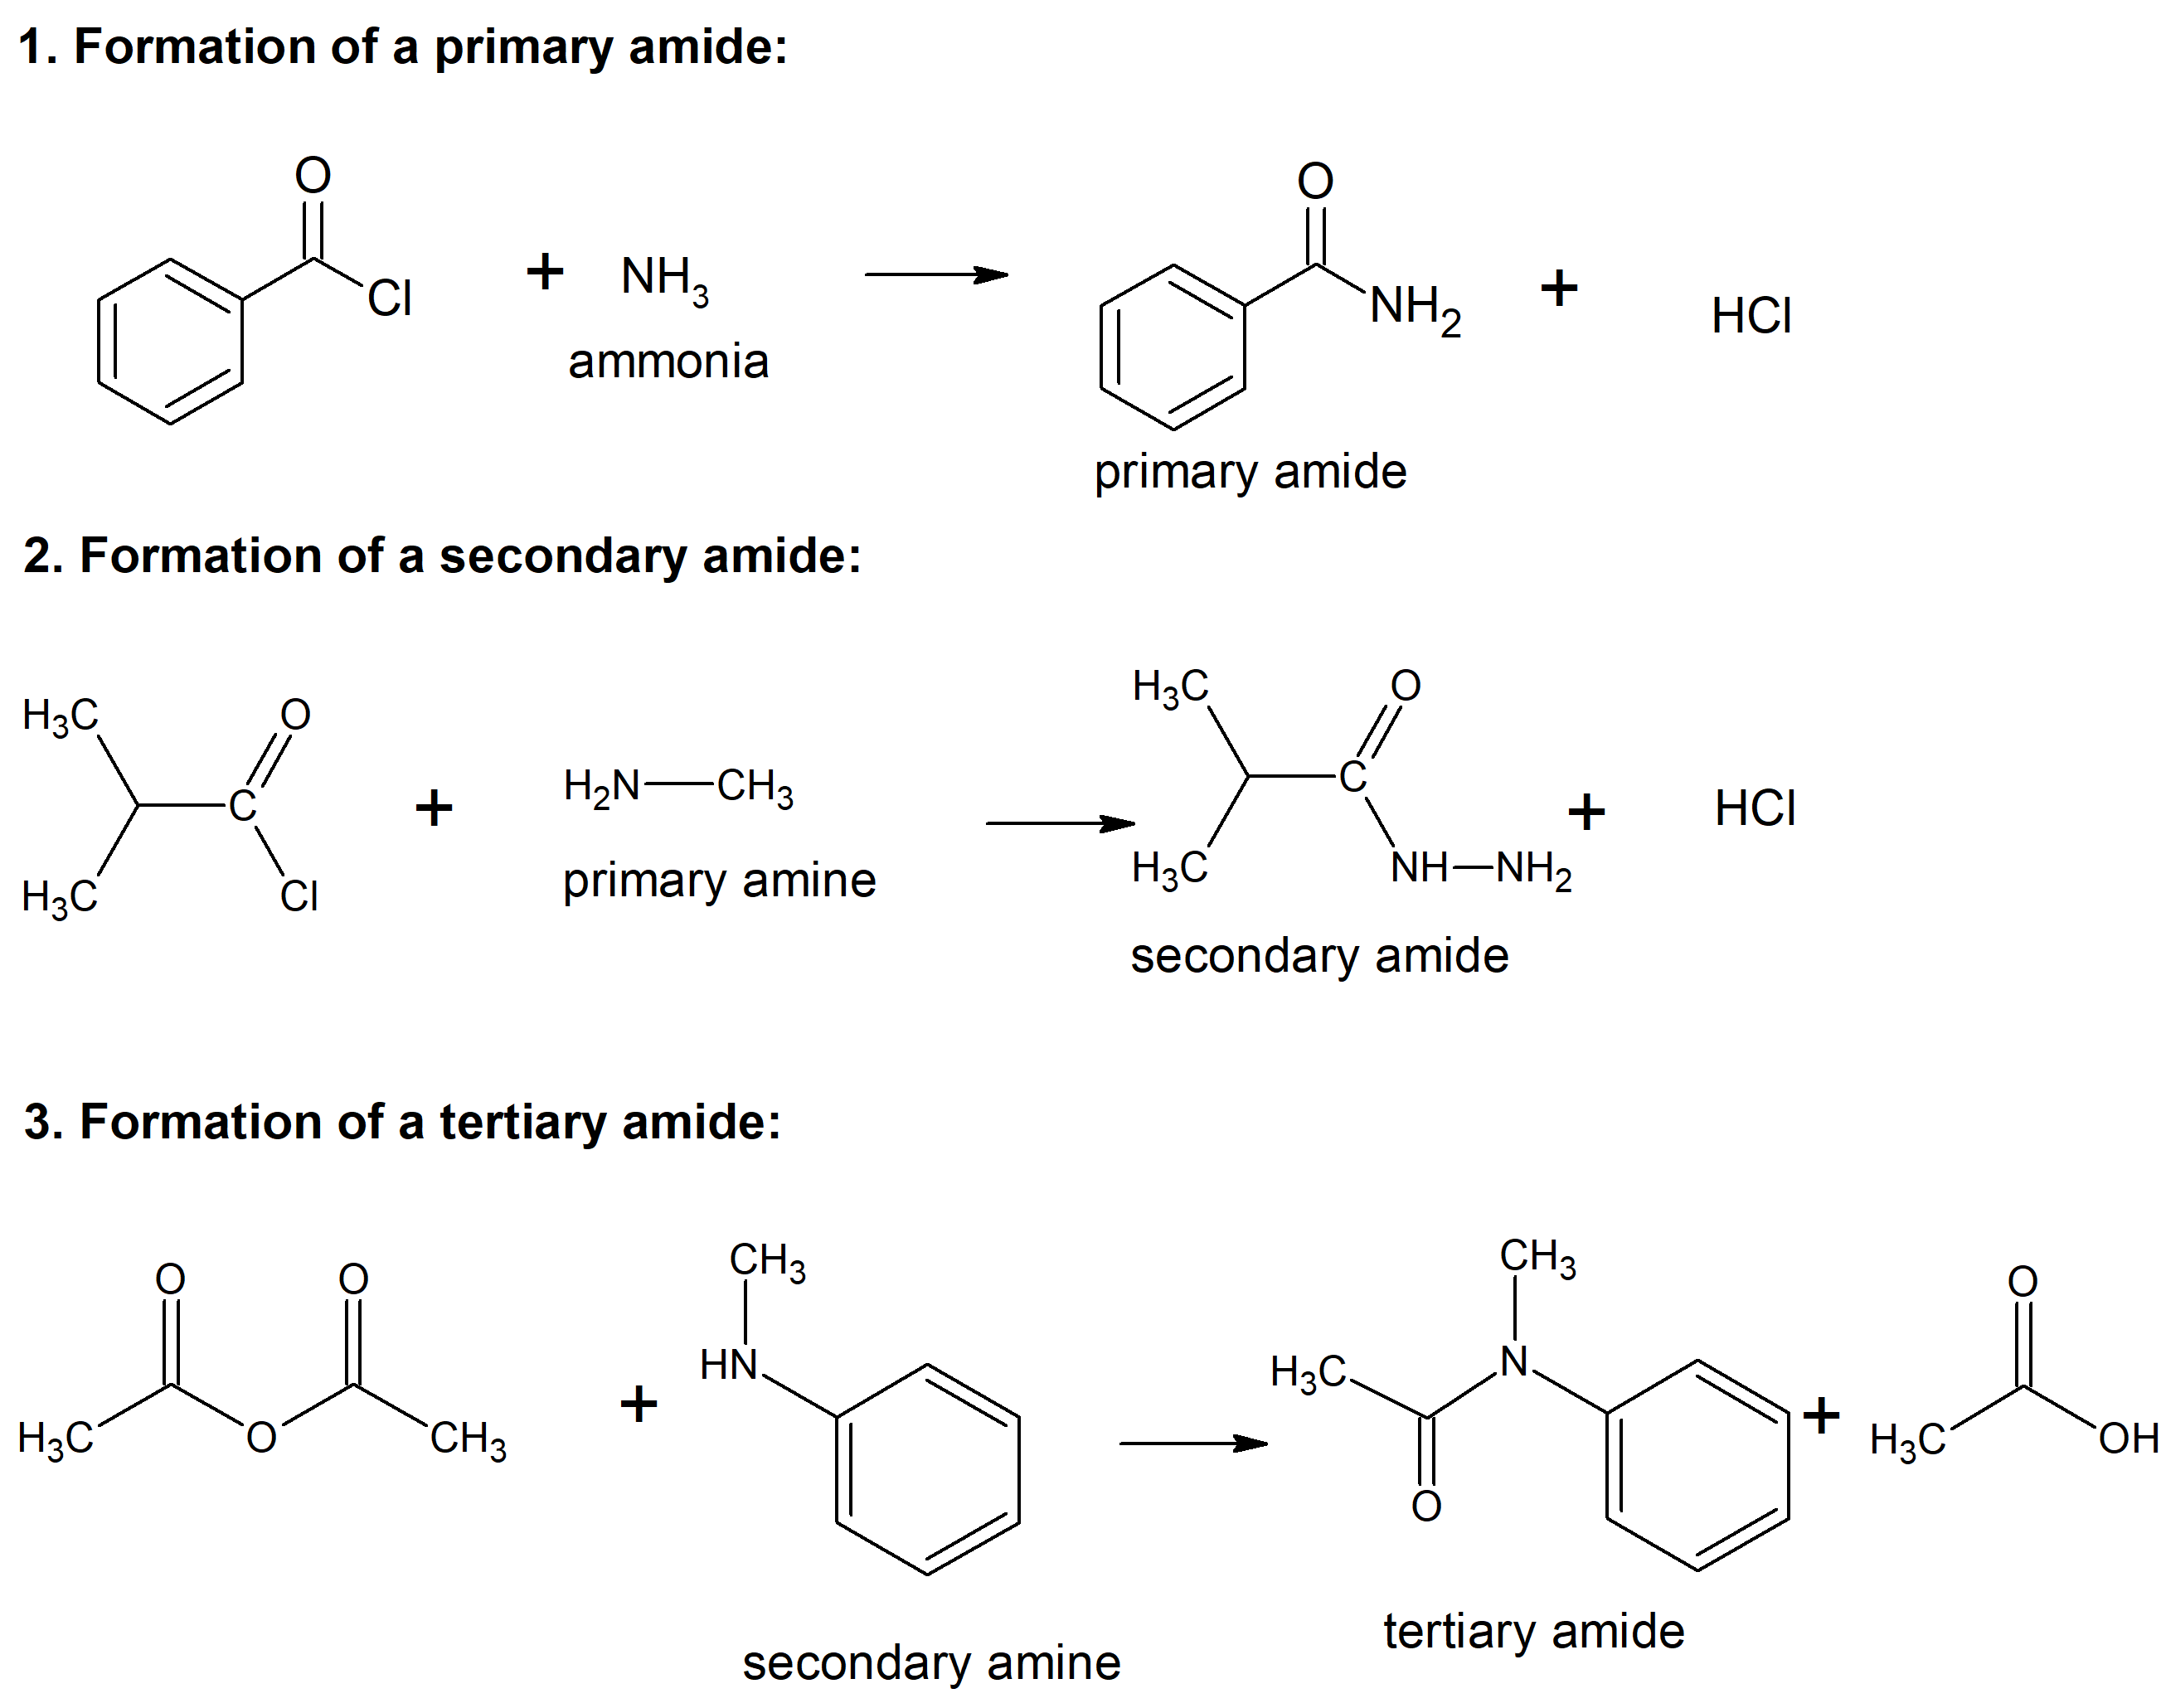
Another area where amides surprise people is their behavior in nucleophilic acyl substitution. Amides are the least reactive carboxylic acid derivative toward nucleophiles. If you have an acid chloride, an anhydride, an ester, and an amide in the same molecule and you add a nucleophile, it will hit the acid chloride first, then the anhydride, then the ester, and leave the amide largely untouched unless you use very harsh conditions. This selectivity is useful but also a trap. I once tried a straightforward aminolysis on an ester in the presence of a benzamide protecting group, assuming the amide would survive. It did survive, but the reaction was so slow that I ended up over-reacting other sensitive groups on the molecule instead of getting clean ester conversion. The workaround was to convert the ester to a more reactive thioester first, which allowed the aminolysis to proceed rapidly at lower temperature while the amide stayed. The Schmidt reaction and Hofmann rearrangement are two named reactions that involve amides and are worth knowing about. The Schmidt reaction uses hydrazoic acid to convert a carboxylic acid or amide into an amine with loss of carbon dioxide. It is reliable but uses HN3, which is toxic and explosive, so it requires proper safety infrastructure. The Hofmann rearrangement converts a primary amide to a primary amine with one fewer carbon atom using bromine and a base. It proceeds through an isocyanate intermediate that is trapped by water. The key pitfall here is that secondary amides do not undergo this rearrangement at all, and if your starting material has any N-substitution, the reaction simply will not work and you will waste time wondering why.
Get the Full Details

Practical Tips That Actually Help
Monitoring amide reactions by TLC can be misleading because amides often have very similar Rf values to their products. I recommend running a quick derivatization test on the TLC plate. Dip the developed plate in a solution of sodium nitrite and sulfuric acid, then heat it. Amides and amines show up as different colored spots, which makes it much easier to track conversion. This simple trick has saved me from running unnecessary extra hours on reactions that were already complete. Purification is another area where amides cause headaches. They tend to tail badly on silica gel because of hydrogen bonding interactions with the silanol groups. Using a small amount of ammonia or triethylamine in your eluent can sharpen the spots significantly. A gradient from 5% to 10% methanol in dichloromethane with 0.1% ammonia works well for most polar amide-containing compounds. Flash chromatography without the base additive usually gives you a smeared band that is impossible to collect cleanly. Amides also pose problems in NMR interpretation. The NH protons can exchange with deuterium if you use CD3OD as your solvent, which makes them disappear from the spectrum. Always use DMSO-d6 or CDCl3 if you need to see those signals. The amide NH in DMSO-d6 typically shows up as a broad singlet between 5.5 and 8.5 ppm depending on substitution and hydrogen bonding. In CDCl3, the signal is often broader and can shift depending on concentration because of intermolecular association.
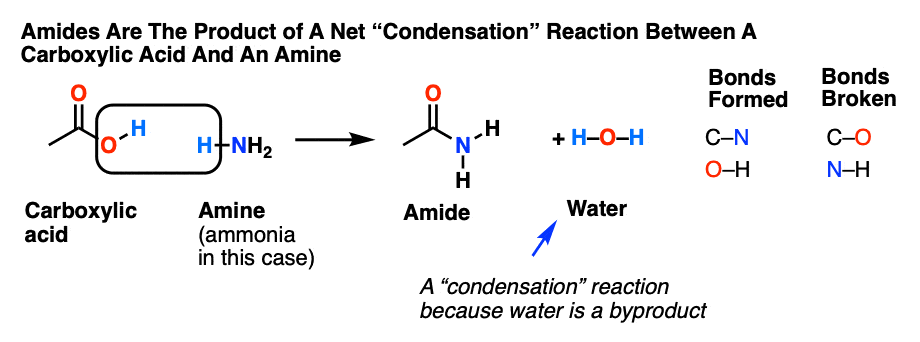
If you are working with sterically hindered amides, expect slower reaction kinetics across the board. N,N-diisopropylamide derivatives, for example, resist hydrolysis and reduction much more than their unsubstituted counterparts. The steric bulk around the nitrogen reduces the resonance stabilization of the amide bond slightly, which might suggest higher reactivity, but in practice the steric hindrance to nucleophilic approach dominates and the reaction becomes very slow. I have seen literature procedures for reducing bulky amides that require 24 hours of reflux even with excess LiAlH4. For large-scale work, amide hydrolysis in concentrated sulfuric acid is safer and more controllable than using hydrochloric acid because there is no gas evolution and the acid can be recovered by dilution and extraction. The trade-off is that sulfuric acid can sulfonate aromatic rings if your substrate has one, so this is not a universal improvement. It depends entirely on your molecule. One final thing that people overlook is the moisture sensitivity of amide activation reagents. If you are using coupling agents like EDC or DCC to form amides from carboxylic acids and amines, the reagents themselves are sensitive to water. EDC hydrolyzes to a harmless urea derivative in the presence of moisture, but it consumes the reagent and lowers your yield. Keeping your solvents anhydrous and your glassware dry is not optional when doing amide bond formation. I lost a whole batch once because I used freshly opened EDC without checking the desiccant in the glovebox, and the humidity had already degraded a significant portion of it. The reaction looked fine by TLC but the isolated yield was half of what I expected based on the conversion.