What S P D Orbitals Actually Are, From Someone Who Has Graded Too Many Student Papers on This Topic
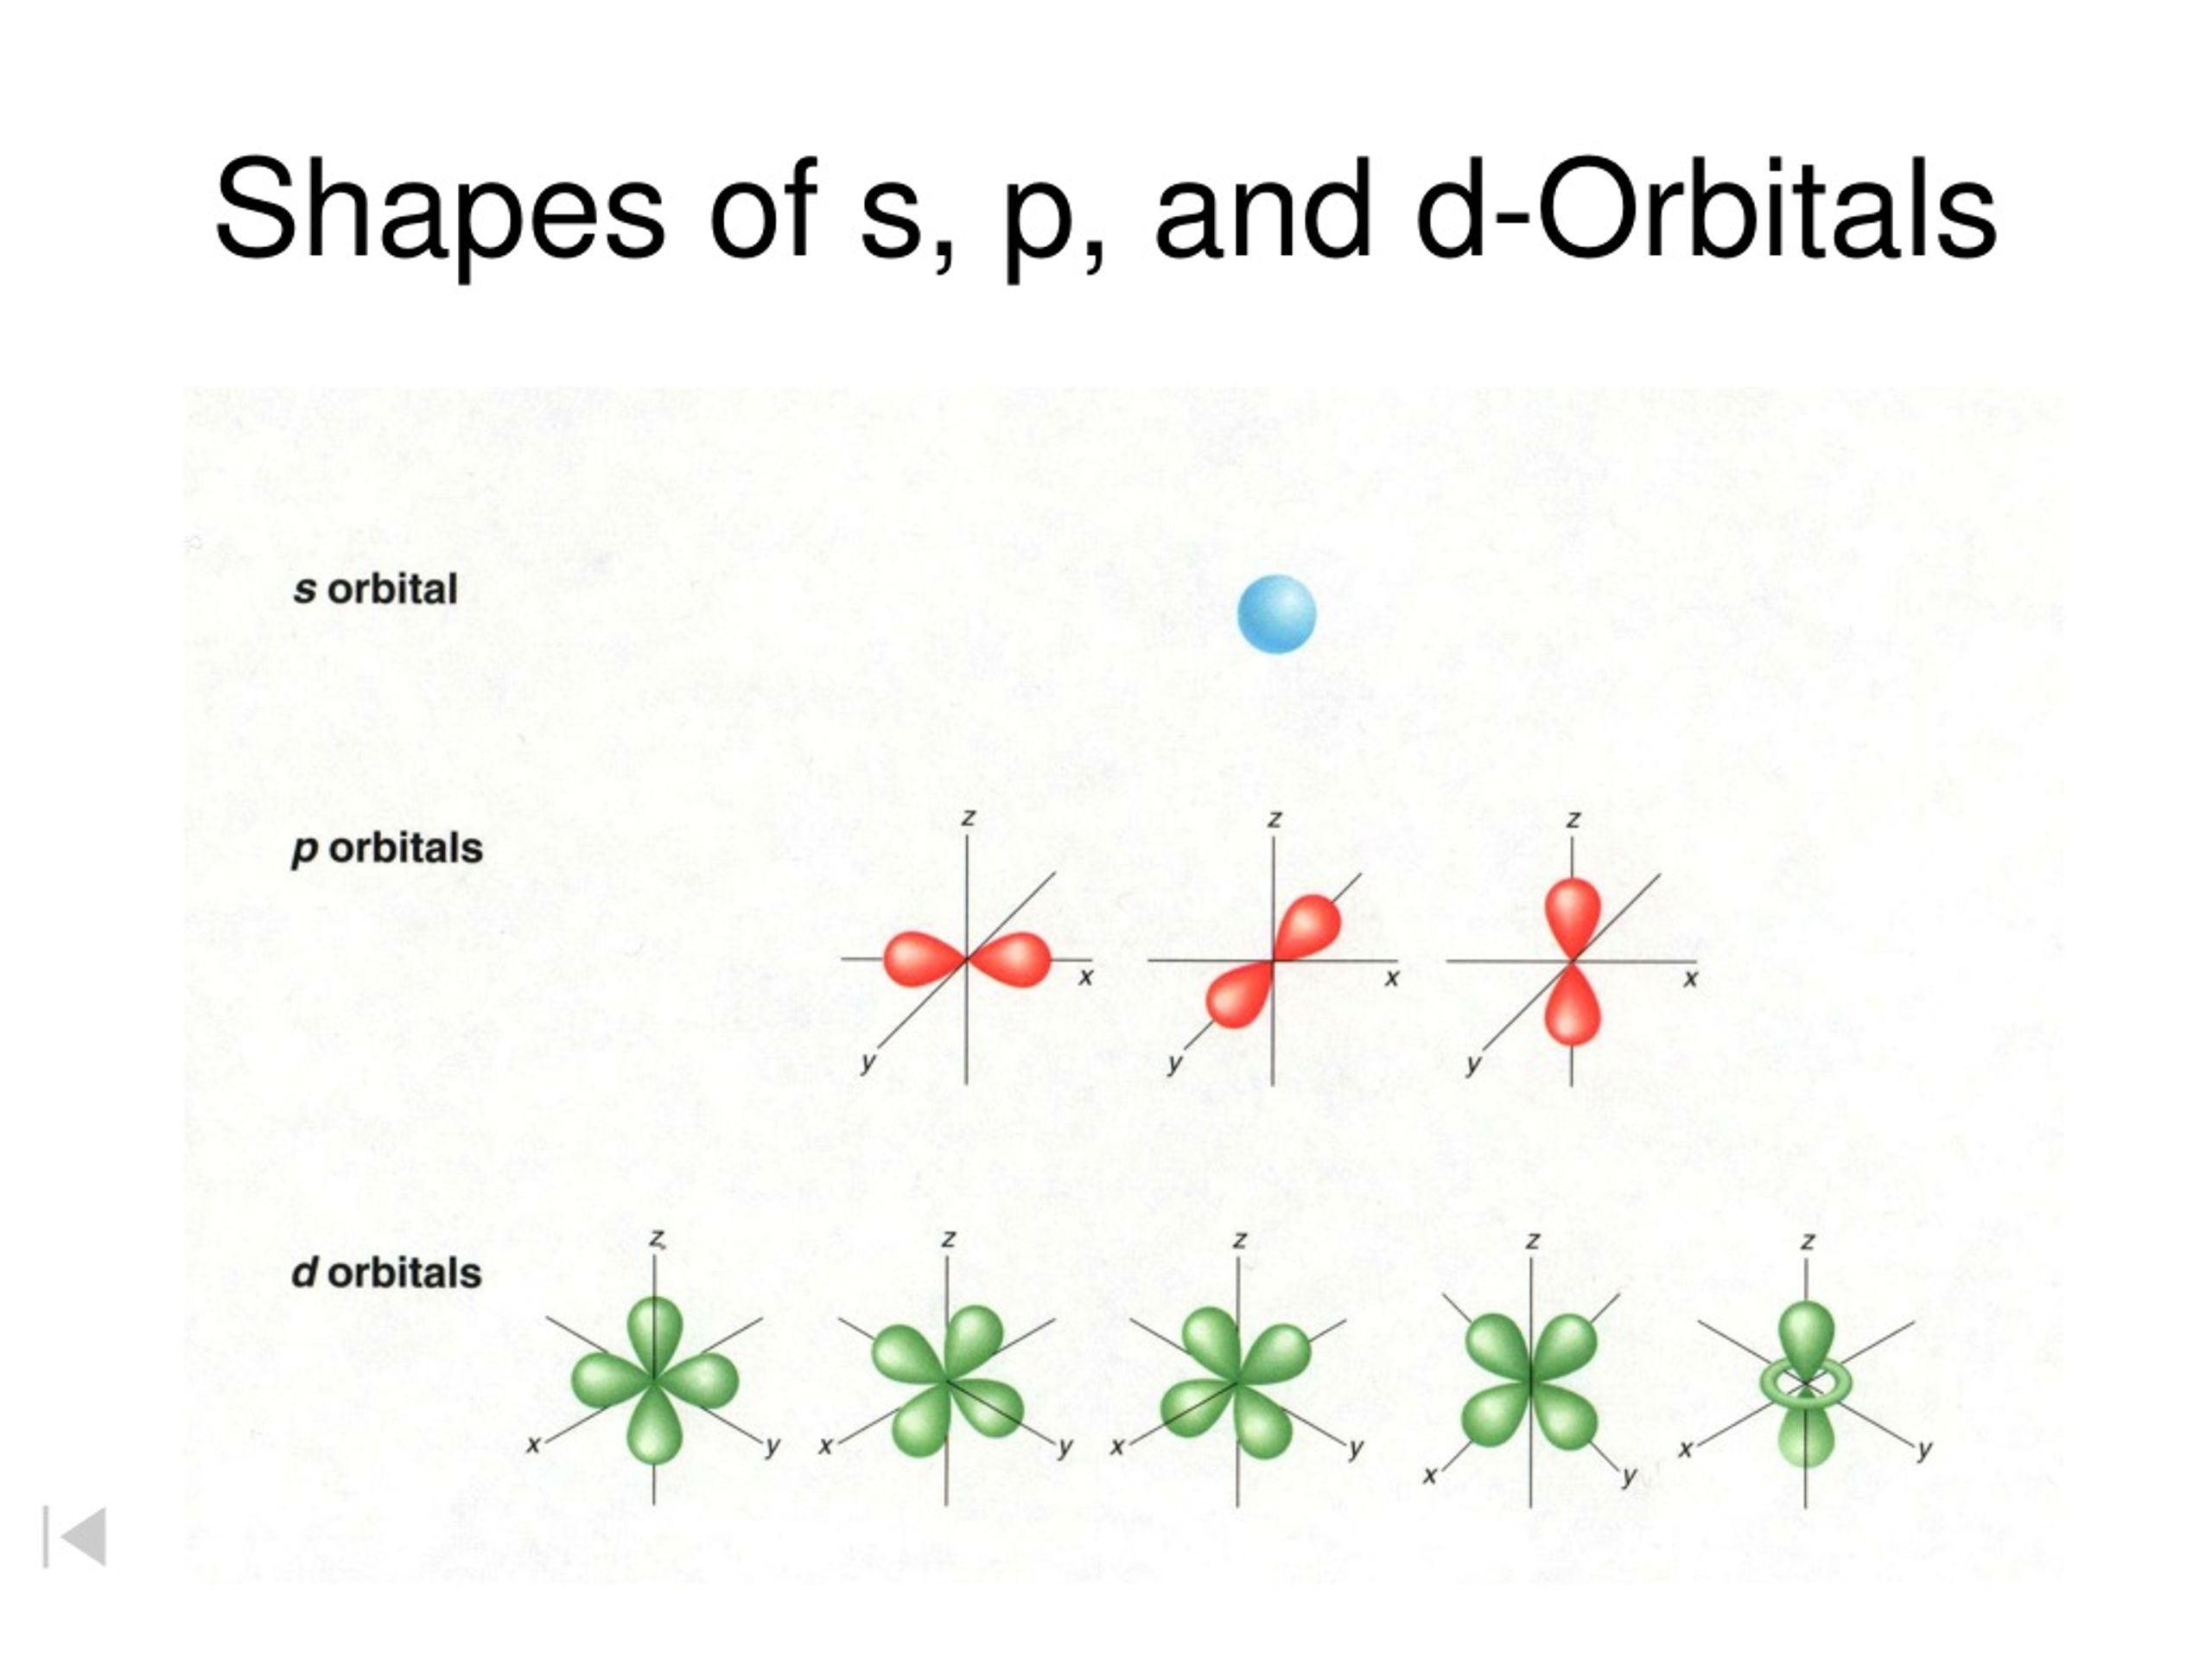
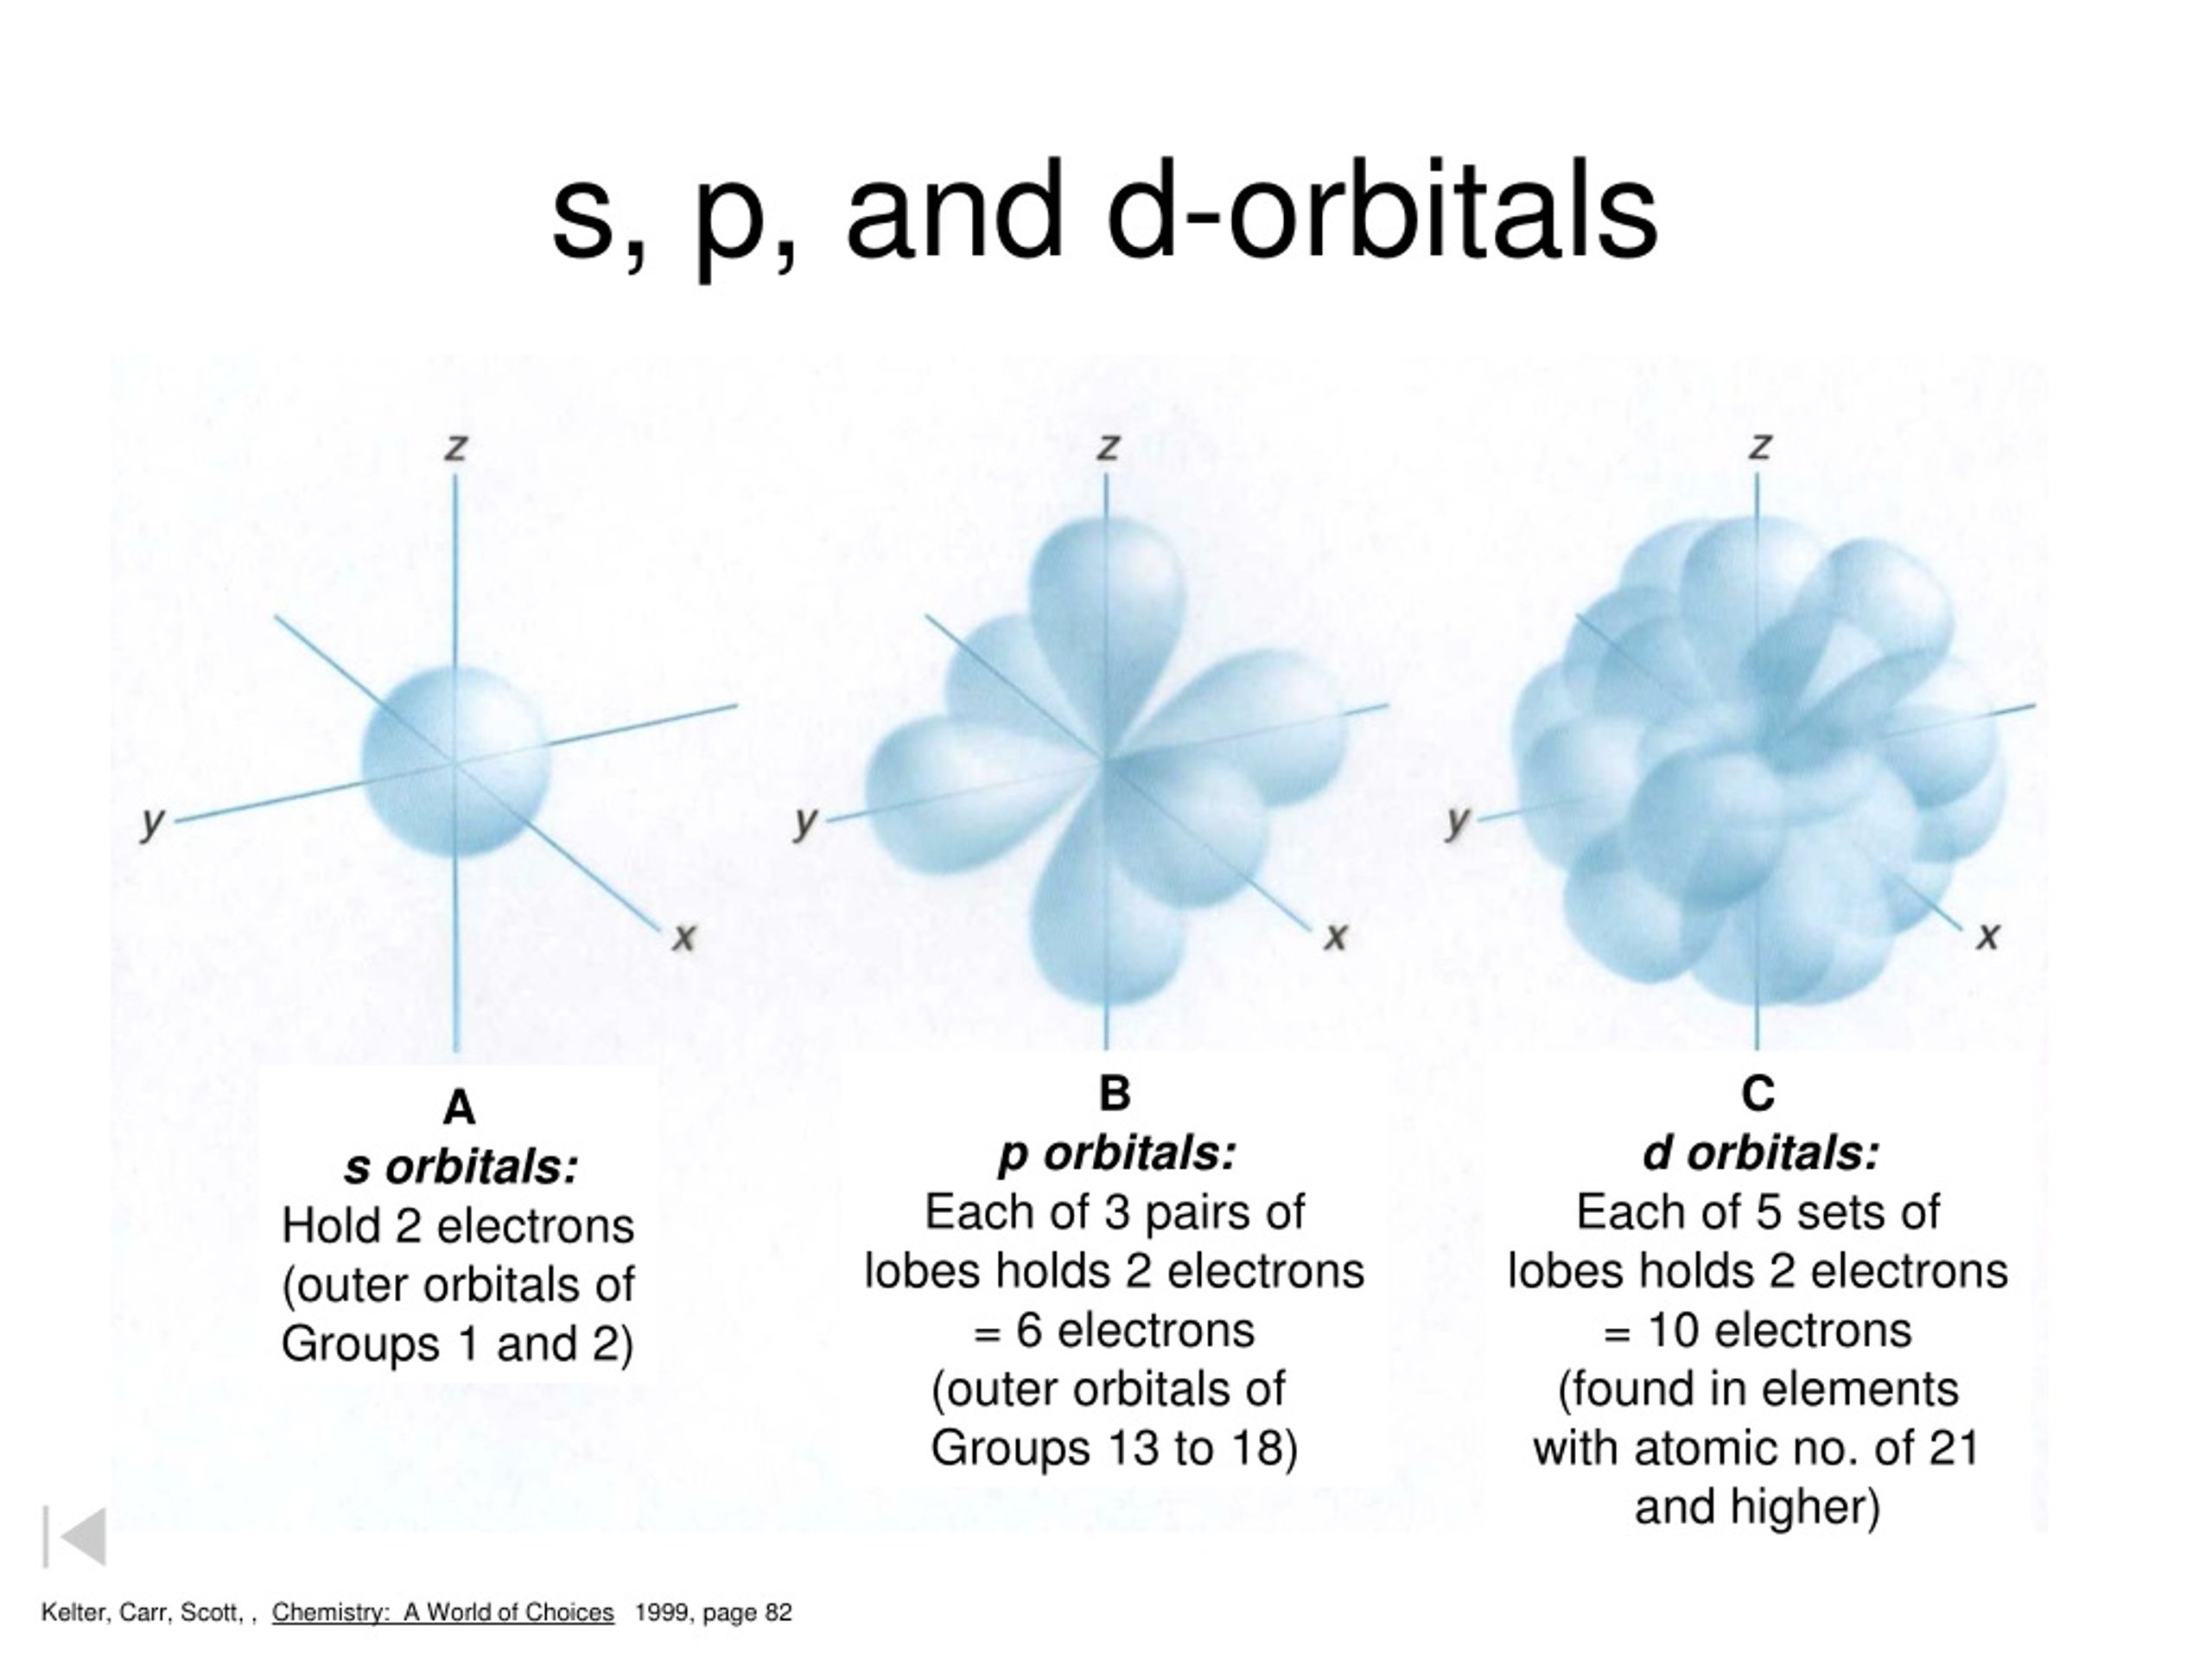
Quantum numbers define where an electron lives. The azimuthal quantum number, l, determines the subshell shape. When l equals zero, one, or two, you get s, p, and d orbitals respectively. This is standard undergraduate physical chemistry. The part people actually struggle with isn't the definition. It's connecting the mathematical description to what you see in a computational output or a lab result. If you are running Gaussian, ORCA, or any similar package and want to understand what the program is actually doing with your electron density, you need to understand how these orbitals behave under different basis sets. Here is the thing most textbooks skip: the shape you draw on a whiteboard is a hydrogenic solution. Real atoms, especially transition metals, don't look like that when you compute them with a proper basis set. The d orbitals on iron in Fe(CO)5 are not the same thing as the d orbitals in a free iron atom. They split. They mix with ligand orbitals. They contract or expand depending on your basis set and your level of theory. I once spent three days trying to debug why my TD-DFT excitation energies were garbage for a platinum complex. The problem wasn't the functional. It wasn't the geometry. It was that I had used a basis set that did not include diffuse functions on the ligands, and the excited state had significant charge-transfer character. Adding diffuse functions to the basis set and re-running changed the lowest excitation energy by about 0.4 eV. That sounds small until you're comparing against experimental UV-Vis data and your error margin was already at 0.3 eV.
The Practical Rules Nobody Emphasizes Enough
Rule one: s, p, and d labels are not rigid categories in a molecule. In a DFT calculation, orbitals are mathematical constructs that emerge from the self-consistent field procedure. An orbital labeled as d-character in one region of space might have significant s or p mixing in another. When you look at an MO diagram from a program output, don't trust the percent composition numbers blindly. Check the basis functions used. Different programs define orbital occupation and characterization slightly differently. Rule two: The ordering of d orbitals depends entirely on symmetry. In an octahedral field, you get t2g and eg. In tetrahedral, it inverts. In lower symmetry, all five can split individually. If you are trying to rationalize magnetic properties or EPR parameters and your crystal field diagram has four d orbitals at the same energy in a system that clearly lacks that symmetry, go back to your geometry and check for distortions. Jahn-Teller effects are annoying but they are not optional. Rule three: For heavy elements, scalar relativistic effects matter more than d-orbital splitting does. If you are modeling anything past the fourth period without a relativistic Hamiltonian or an effective core potential, your d-orbital energies are qualitatively wrong. Not slightly off. Wrong. The contraction of s and p orbitals under relativistic conditions indirectly expands and destabilizes d orbitals, and this changes everything from bond lengths to oxidation state stability. I learned this the hard way when my calculated reduction potential for a gold complex was off by more than a volt because I had used a non-relativistic all-electron basis set on gold. Switching to a relativistic ECP fixed it immediately.
When S P D Orbitals Break Down Completely
Multi-reference systems are where single-determinant orbital pictures fail. Transition metal clusters, bond-breaking scenarios, and certain excited states cannot be described adequately by one set of s, p, and d orbitals. CASSCF or DMRG methods exist for these cases, but they require you to pick an active space. Picking the wrong active space is worse than using DFT with a decent functional, because at least DFT gives you a physically reasonable answer most of the time. Wrong active space gives you a beautifully converged calculation that answers the wrong question. I have seen postdocs spend weeks on this mistake. Start small. Put the metal d orbitals and the directly bonding ligand orbitals in your active space. Nothing else. Add more later if diagnostics tell you to. The s, p, d orbital framework is a useful language for discussing electronic structure. It is not the territory. Treat it like one.
Get the Full Details