SN2 Mechanisms Don't Care About Your Textbook Diagrams
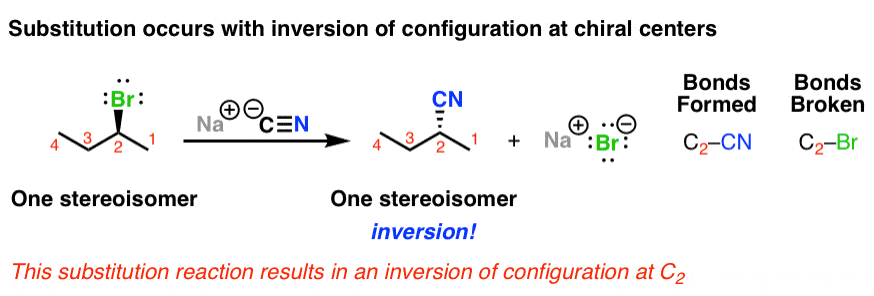
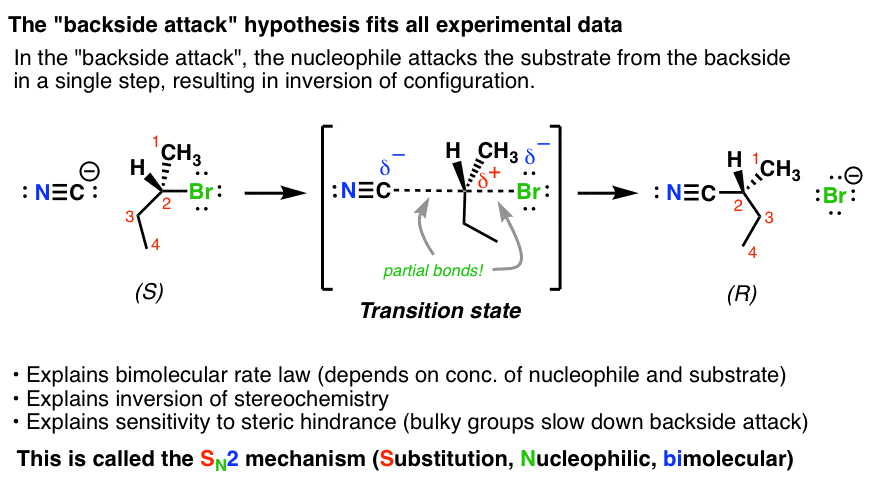
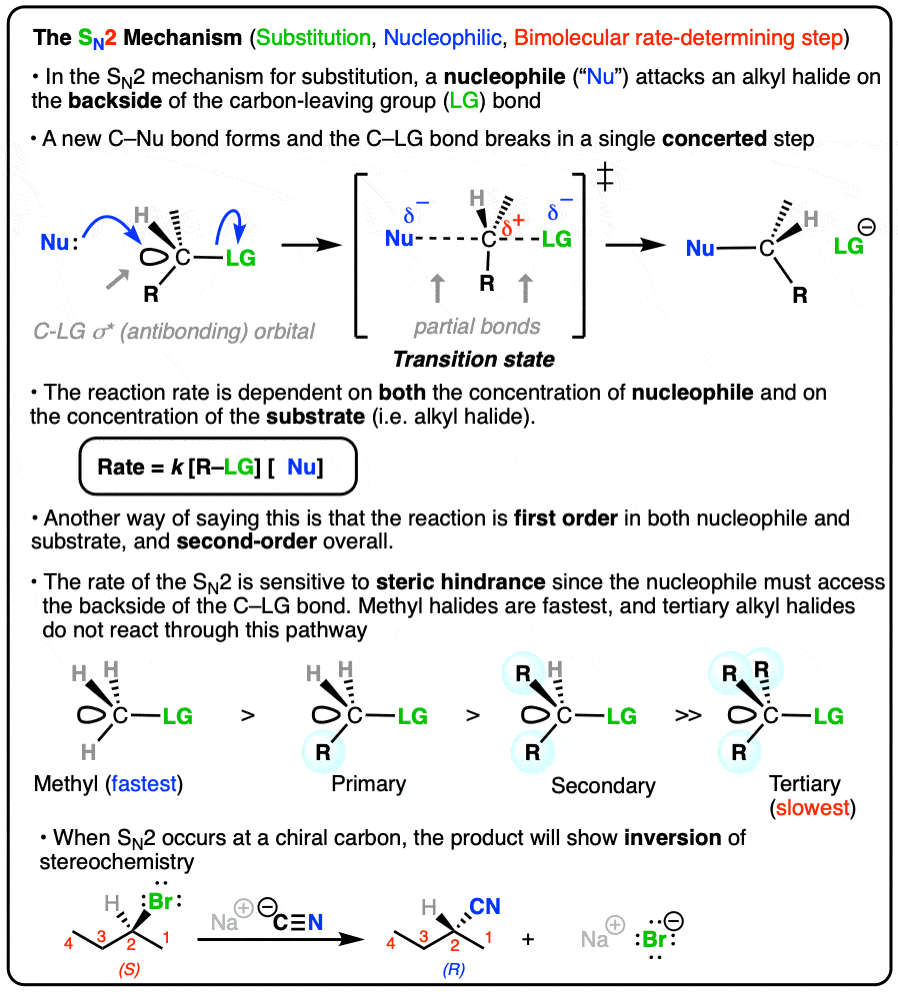
The Sn2 Reaction Organic Chemistry gets explained as a clean, single-step backside attack with perfect stereochemical inversion. That's true if you're drawing it on paper and nobody's watching. In practice it's messier, and the mess is where you lose points or waste a day in the fume hood. Let's get the mechanics out of the way first, because everyone who reads this needs to internalize the orbital picture before anything else. The nucleophile donates a lone pair into the * antibonding orbital of the C–LG bond. The transition state has five groups around carbon, all roughly equidistant. The leaving group departs as the nucleophile bonds. Inversion happens because the nucleophile has to approach from the opposite face of the leaving group. That's it. Backside attack, concerted, bimolecular kinetics. Rate = k[nucleophile][substrate]. Done with the definitions. Now the stuff that actually matters for getting this right in a lab or on an exam that isn't trying to trick you (which is rarer than you'd hope).
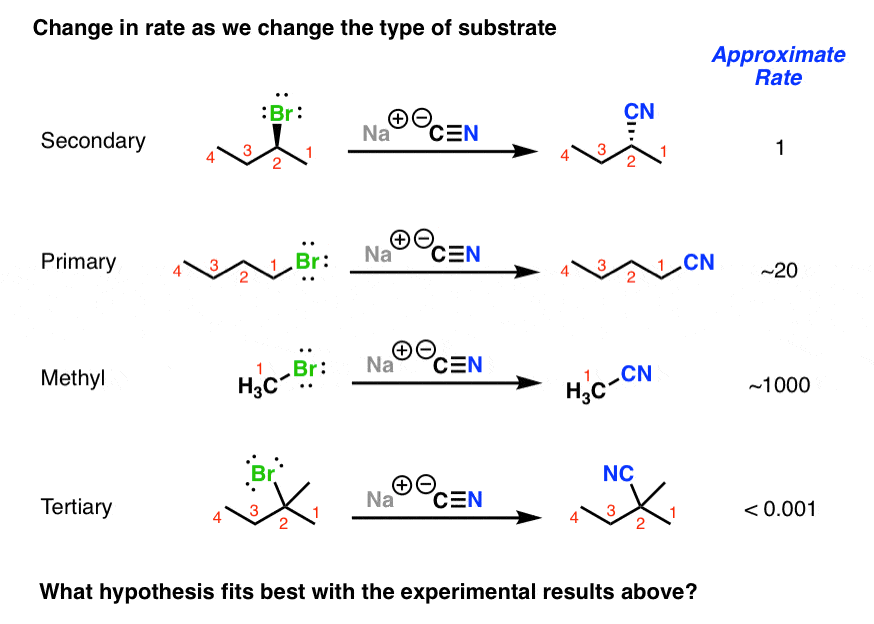
SN2 is extremely sensitive to steric environment at the carbon bearing the leaving group. Methyl and primary substrates are fast. Secondary substrates are slow and competitive with elimination. Tertiary substrates don't do SN2 at all under normal conditions, period. I can't stress that enough. When I was a grad student, a postdoc kept giving me secondary tosylates in polar aprotic solvents and expecting clean displacement. The reaction worked, but so did E2, and the product ratio depended on things he wasn't tracking: the exact hydration of the solvent, the concentration of base impurities in the nucleophile salt, and how long the substrate had been sitting in the bottle. By the time I flagged it, we'd wasted three weeks of material. The fix was simple but annoying. Switch to a primary substrate whenever possible. If you can't, use a non-basic nucleophile like an azide or thiolate, keep the temperature low, and verify the leaving group purity before running anything. Here's a counter-intuitive point that textbooks barely touch: the solvent effect is not just about solvation of the nucleophile. Yes, polar aprotic solvents like DMSO, DMF, and acetonitrile accelerate SN2 by failing to hydrogen-bond to the nucleophile, which raises its ground-state energy and makes it more reactive. But these same solvents also stabilize the charged transition state through dipole interactions. The rate enhancement isn't purely an entropy thing. It's a combination of raising nucleophile energy and stabilizing the TS. In water, the nucleophile is heavily solvated, which drops its energy and slows the reaction, but water also stabilizes the TS well. The net effect is that SN2 is usually slower in protic solvents, but the gap isn't as dramatic as people think for charged nucleophiles. Another thing beginners consistently miss: the leaving group matters more than the nucleophile for determining whether SN2 is even viable. A poor leaving group like fluoride or hydroxide basically shuts down the reaction regardless of how good your nucleophile is. Good leaving groups are weak bases, and the trend tracks with pKa of the conjugate acid. Tosylate, mesylate, triflate, iodide, bromide are your reliable options. Chloride is borderline and needs either a really good nucleophile or an activating additive. I once tried an SN2 with a chloride substrate and a strong alkoxide nucleophile in THF at reflux for 18 hours and got basically nothing. Switched the substrate to the corresponding tosylate, same conditions, reaction finished in about 40 minutes. The nucleophile didn't change. The leaving group did. That's the lesson.
Watch out for neighboring group participation, which can completely fake out your stereochemical analysis. If there's a nearby heteroatom with a lone pair, it can act as an internal nucleophile, form a cyclic intermediate, and then get attacked by your external nucleophile. The net result can be retention of configuration instead of inversion, which looks like the SN2 mechanism failed until you realize two SN2 steps happened in sequence. I ran into this with a -hydroxy tosylate and had to figure out why my NMR showed retention instead of the expected inversion. Took a day to work through it, but the second-order kinetics and the isotopic labeling experiment confirmed the neighboring group pathway. The workaround is to protect the neighboring hydroxyl as an ether before running the displacement, or to accept the neighboring group participation if it gives you the stereochemistry you actually want. For the practical execution: use a polar aprotic solvent, keep your nucleophile concentration high relative to the substrate to favor SN2 over E2, maintain moderate temperatures to keep elimination in check, and make sure your substrate is genuinely primary or methyl if you want a straightforward reaction. Secondary substrates require more optimization and you should expect competing pathways. Tertiary substrates need a different mechanism entirely, usually SN1 or E1 depending on conditions. If you need a reference, the classic March's Advanced Organic Chemistry covers the kinetic and stereochemical details comprehensively. For a hands-on perspective, the Vogel textbook has procedures that show the real yield variation you get from solvent purity and leaving group quality.