Getting the Two Concepts Straight Before You Waste Afternoon Time
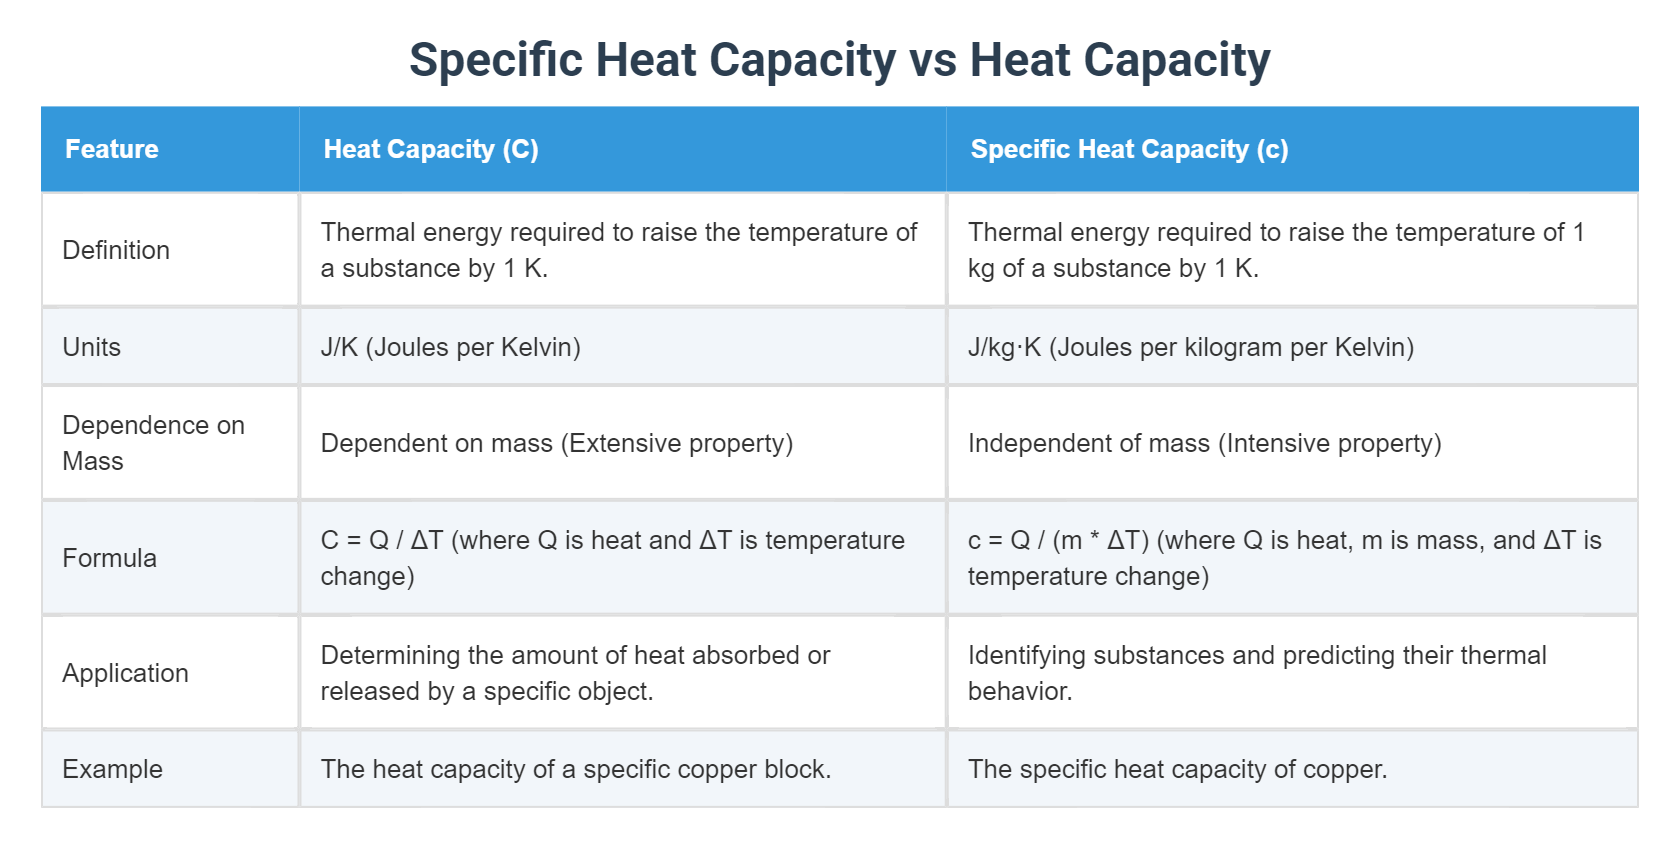
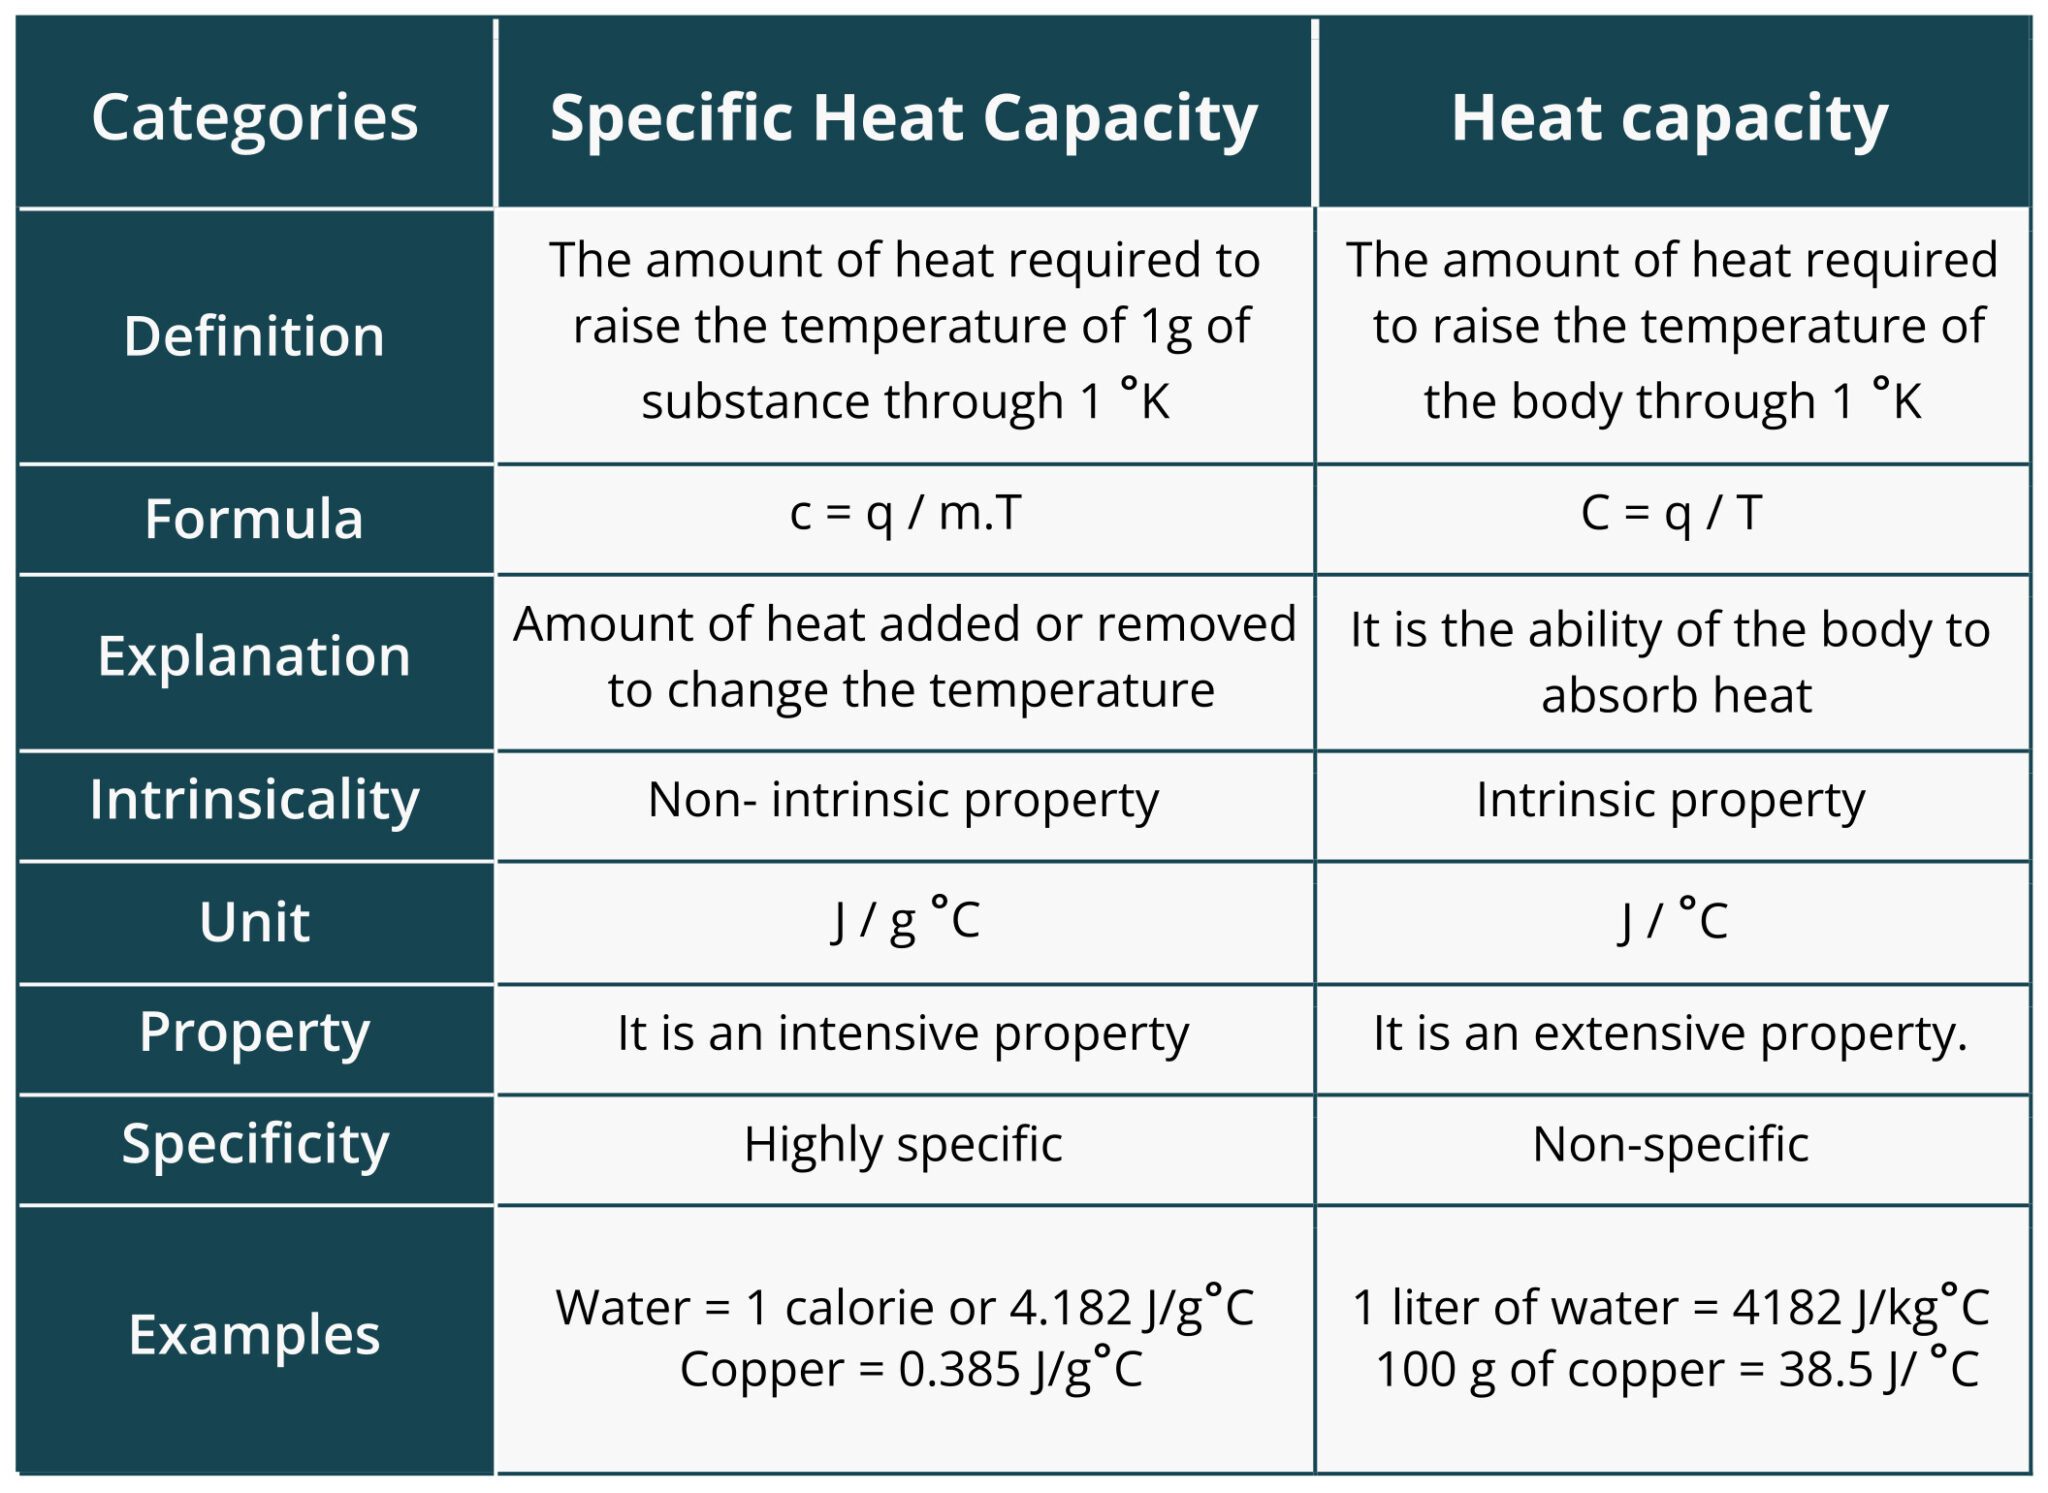
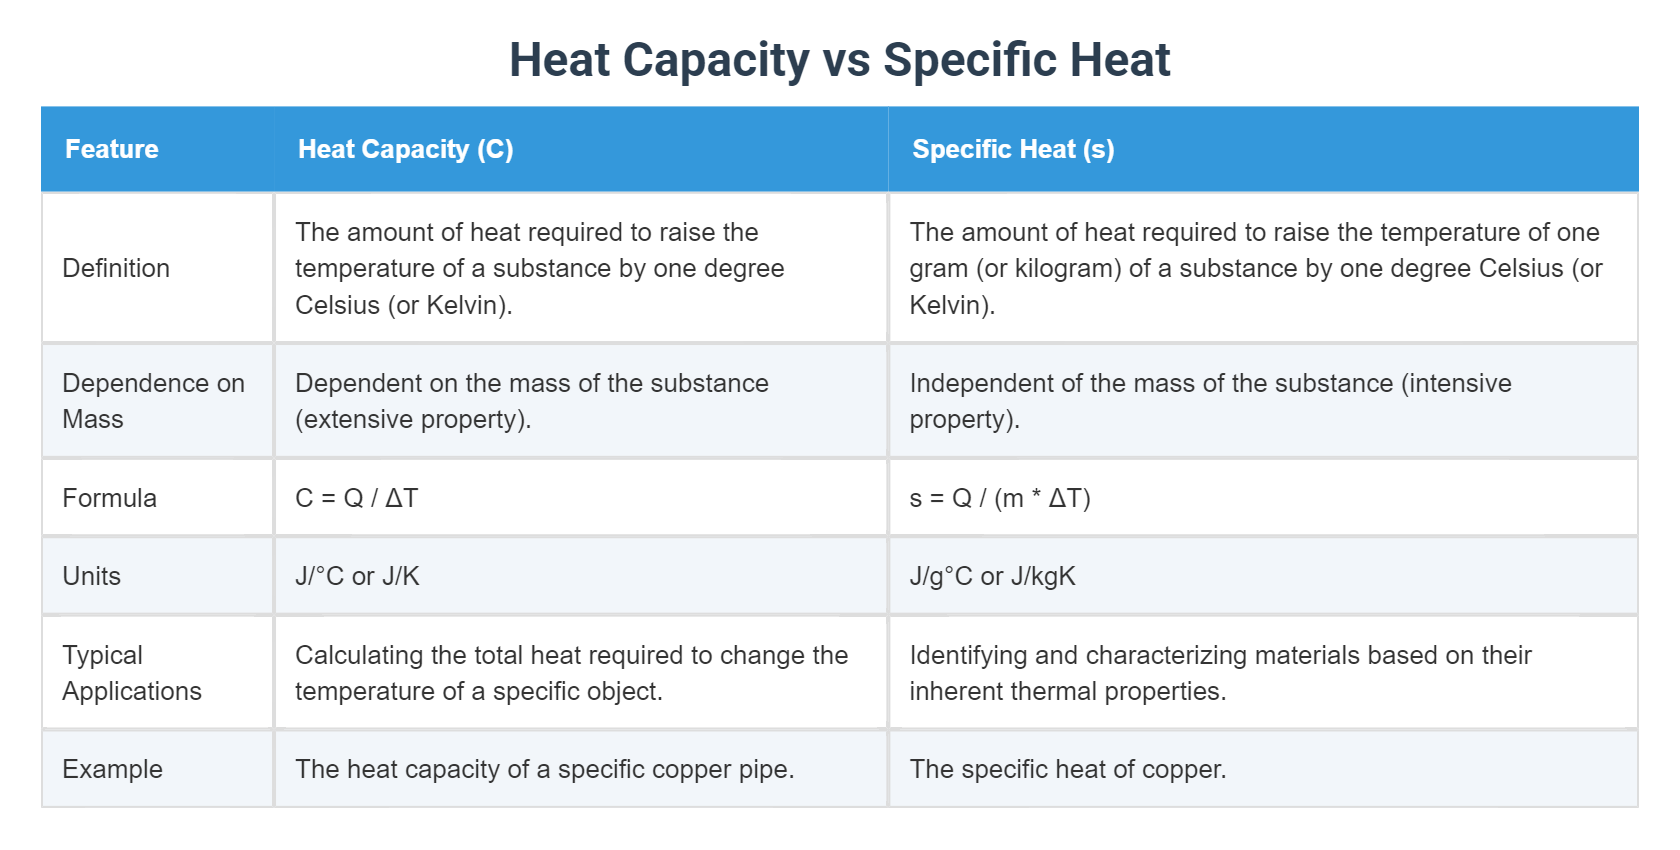
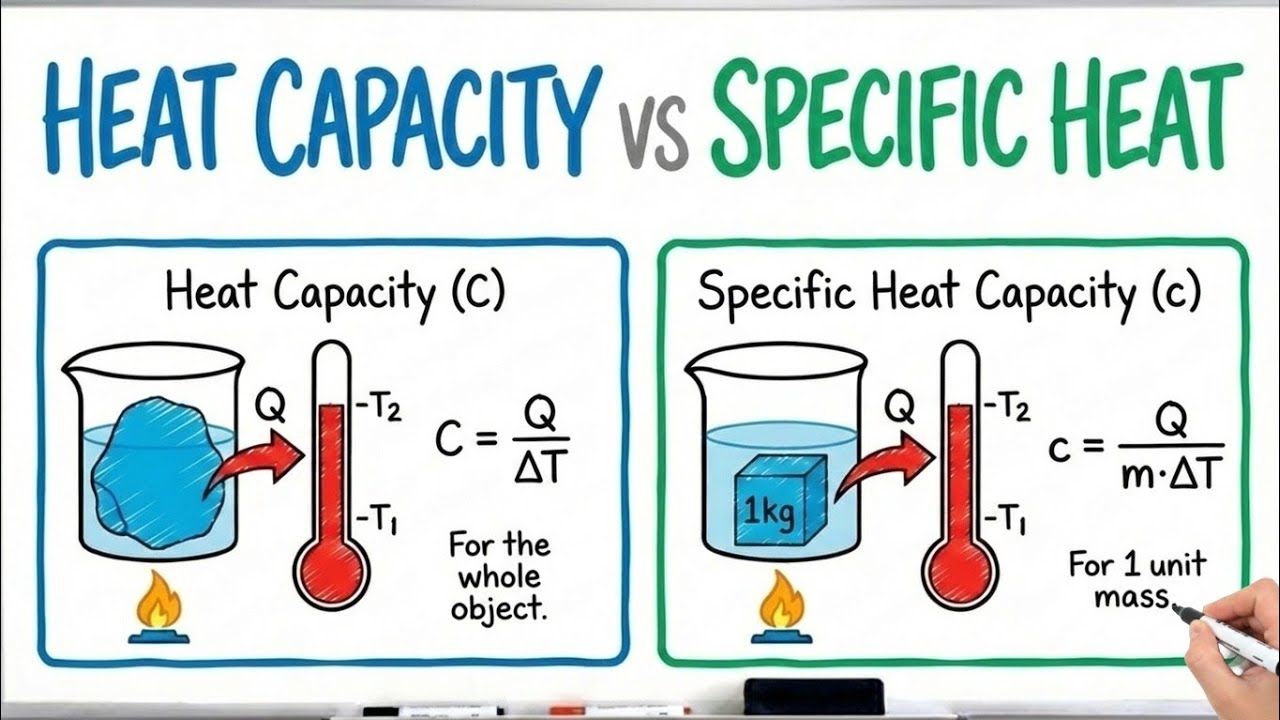
I've seen engineers mess this up on real projects more times than I can count. The confusion isn't particularly deep, but it's expensive when it happens. I need to get into why these two things matter separately, how to actually use them in calculation, and what goes wrong when you treat them as interchangeable. Heat capacity is the amount of energy needed to raise the temperature of an entire object by one degree. Specific heat is the amount needed to raise one unit mass of a substance by one degree. That's the textbook version. The real version is that specific heat is a property of the material itself, while heat capacity is a property of your particular sample. A gallon of water and a teaspoon of water have wildly different heat capacities but the same specific heat. I've been doing thermal analysis for a while now, and the most common mistake is grabbing a specific heat value from a table and multiplying it by mass without checking whether the value is quoted per gram, per mole, or per kilogram. The numerical result can be off by factors of 1000 depending on which convention your source uses. Always check the units. Once. Before you start building anything around the number.
The relationship is straightforward if you keep it straight: C = m × c, where C is heat capacity, m is mass, and c is specific heat. That's it. It's not glamorous. It's also enough to get you in trouble if you're sloppy about units or if you assume the specific heat is constant across temperature ranges where it isn't. Here's a thing most introductory courses gloss over: specific heat changes with temperature. For water it's relatively stable over normal lab ranges, maybe two or three percent between 10 and 40°C, so you rarely notice. For metals and gases it's a different story. Aluminum's specific heat nearly doubles between room temperature and 500°C. If you're doing anything involving phase change or high temperature without accounting for that variation, your numbers are going to drift. I ran into this on a heat exchanger project a few years back. We were sizing a thermal buffer for a process that cycled between 25°C and 350°C, and we had used a constant specific heat value from a reference table at room temperature. The calculated buffer mass was about 30% too small. We caught it during the prototype testing phase when the system couldn't hold temperature through the cycle. The fix was pulling a temperature-dependent Cp curve for the aluminum alloy and integrating it over the operating range rather than using a single value. It added maybe an hour of work upfront and saved us from a redesign that would have taken weeks. Another practical detail that bites people: heat capacity measurements differ depending on whether you hold pressure constant or volume constant. For liquids and solids the difference is usually tiny because those materials don't expand much. For gases it matters enormously. Cp is always larger than Cv for a gas, and the gap is roughly the gas constant R for ideal gases. If you're working with pressurized gas systems or combustion calculations and you grab the wrong one, you won't just be slightly off. You'll be significantly off. I once saw a simulation fail because someone used Cp for air in a constant-volume combustion model. The peak temperature came out about 200K too low, and nobody noticed until someone actually built the thing and it didn't perform like the model predicted.
When I need to measure heat capacity myself, I usually go with a simple differential scanning approach. You heat a known mass of the material at a controlled rate, record the temperature rise, and back out the heat capacity from the power input. The equipment isn't exotic. A decent thermometer, a way to deliver known energy, and data logging. The trick is making sure your heat losses are small compared to the signal. If you're measuring something with low heat capacity in a drafty room, your data will be noisy. I typically insulate the sample chamber and do a blank run with no sample to characterize the background loss rate, then subtract that from the measurement. It cuts the uncertainty from maybe 15% down to around 3% for most common materials. For quick estimates in the field I sometimes use a mixing method instead. You heat a sample to a known temperature, drop it into a measured mass of water at a known lower temperature, and measure the equilibrium point. It's old-school but it works well enough for materials with moderate heat capacity. The limitation is that it assumes no heat escapes to the environment during the mixing, which is never strictly true. You can improve it by doing the mix in an insulated container and waiting for a stable reading rather than chasing the first equilibrium point. The first reading is almost always biased low because the thermometer hasn't equilibrated yet. Here's something else people miss: phase transitions. During a phase change, the effective heat capacity becomes effectively infinite because you're dumping energy into the material and the temperature doesn't budge. The energy goes into the latent heat, not the sensible heat. If you're running a simulation that includes melting or boiling and you don't account for the latent heat term separately, the model will either predict the wrong temperature or reach the phase transition point and then continue heating as if nothing is happening. I've seen both errors in student projects and in industrial settings where the person running the model didn't really understand what they were modeling. The workaround is straightforward but easy to overlook: add a step function for the latent heat at the transition temperature, or use a material library that already includes it.
Get the Full Details
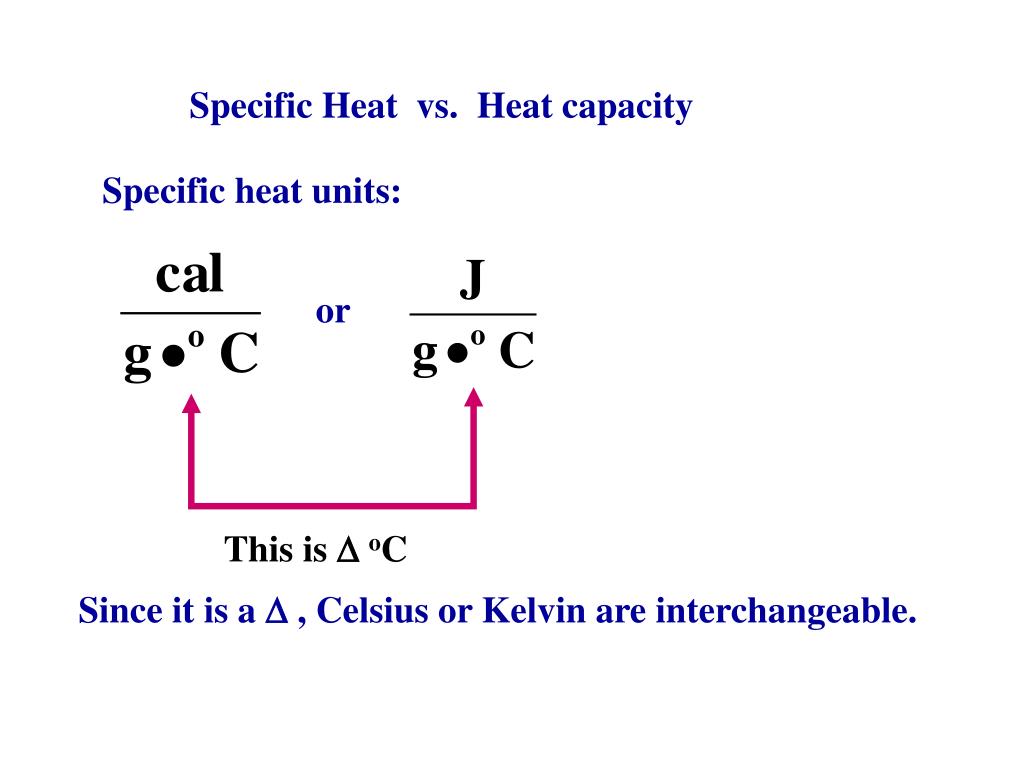
Specific heat data can also vary with purity and microstructure. An alloy's specific heat isn't always a simple weighted average of its components. Impurities, grain boundaries, and dislocation density all affect the vibrational modes that store thermal energy. For most engineering work you don't need to worry about this, but if you're working with precision instrumentation or novel materials, the tabulated values might not match your actual sample by a noticeable amount. I learned this the hard way when characterizing a sintered ceramic for a thermal barrier coating. The vendor's Cp data for the powder didn't match what we measured in the sintered part, and the difference was enough to throw off our thermal cycling predictions. X-ray diffraction showed the sintered part had a slightly different phase composition, which shifted the heat capacity. You can't always rely on the handbook when the material has been processed. One last practical note on units that causes actual problems: the calorie, the joule, the BTU. Different fields use different energy units and different mass or mole bases. Your specific heat might come in J/g·K, kJ/kg·K, cal/g·°C, or BTU/lb·°F, and the numerical values are all different even though they describe the same physical quantity. Converting between them is trivial math, but doing it wrong while tired at 11pm on a deadline is how mistakes get into design documents. I keep a conversion table on my desk now and I check it every time I switch between unit systems. It saves arguments with colleagues and it saves rework.