Getting Your Spectrophotometric Iron Results Right
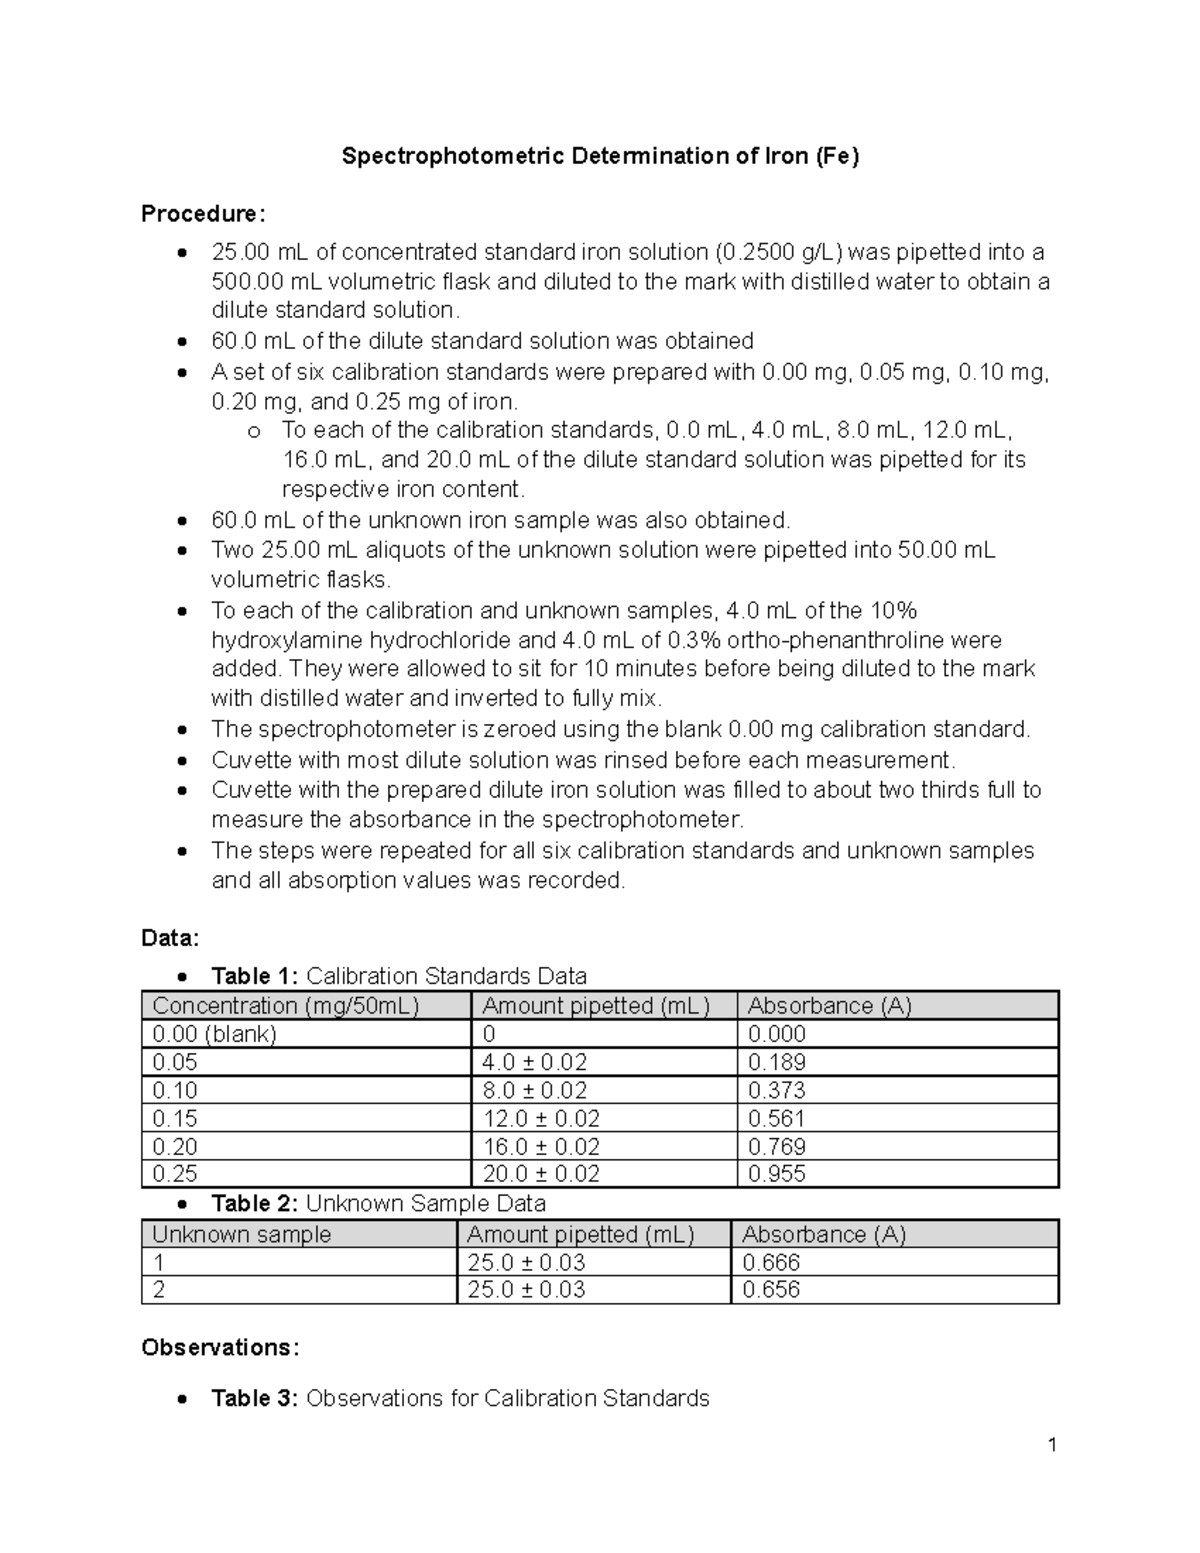
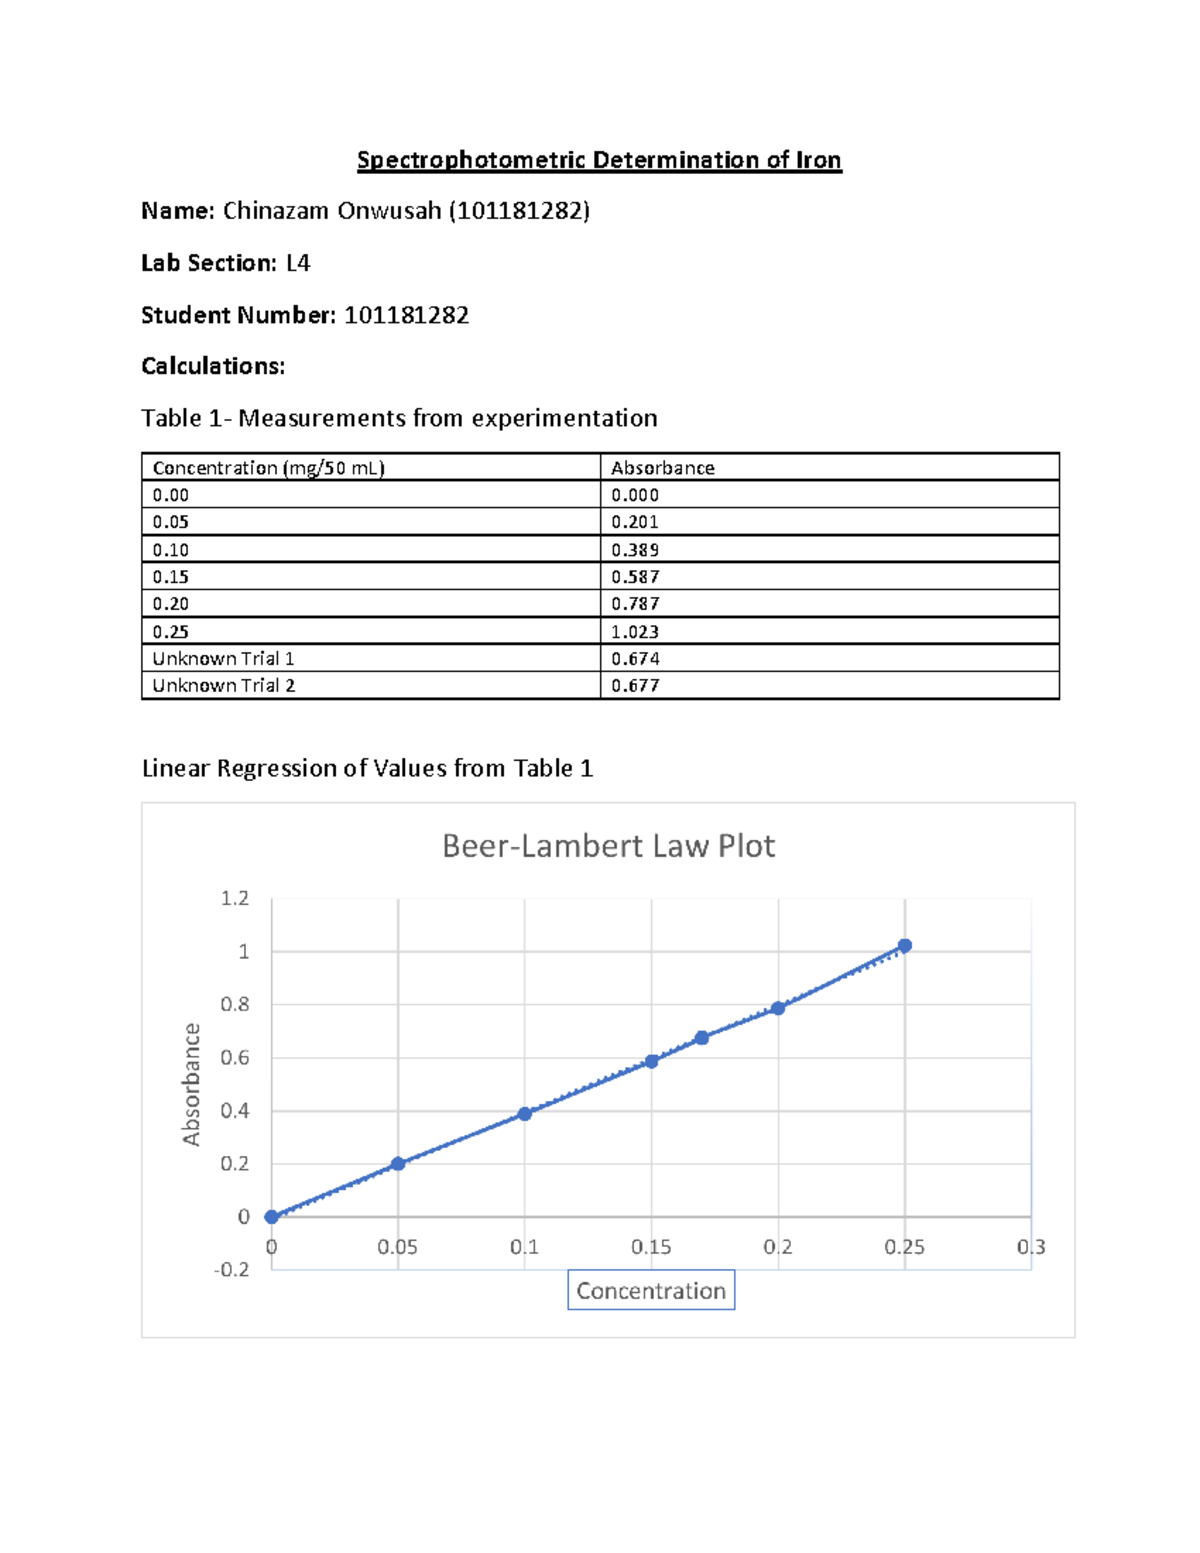
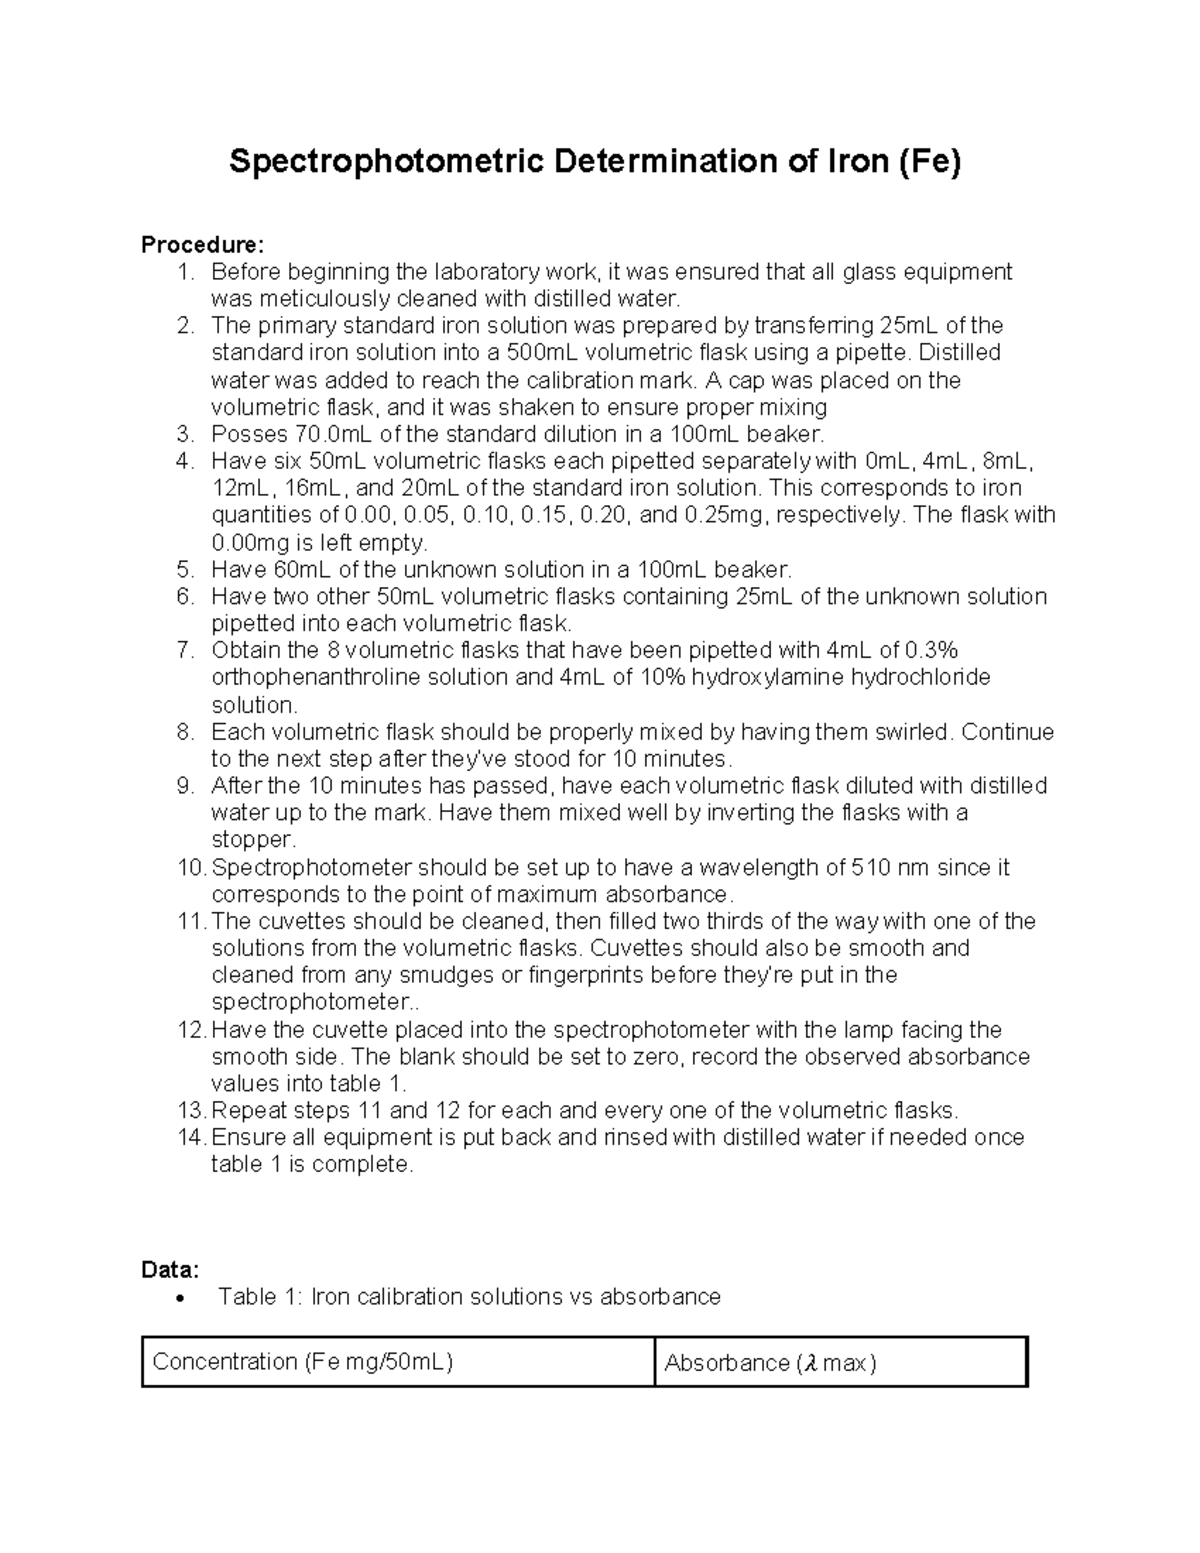
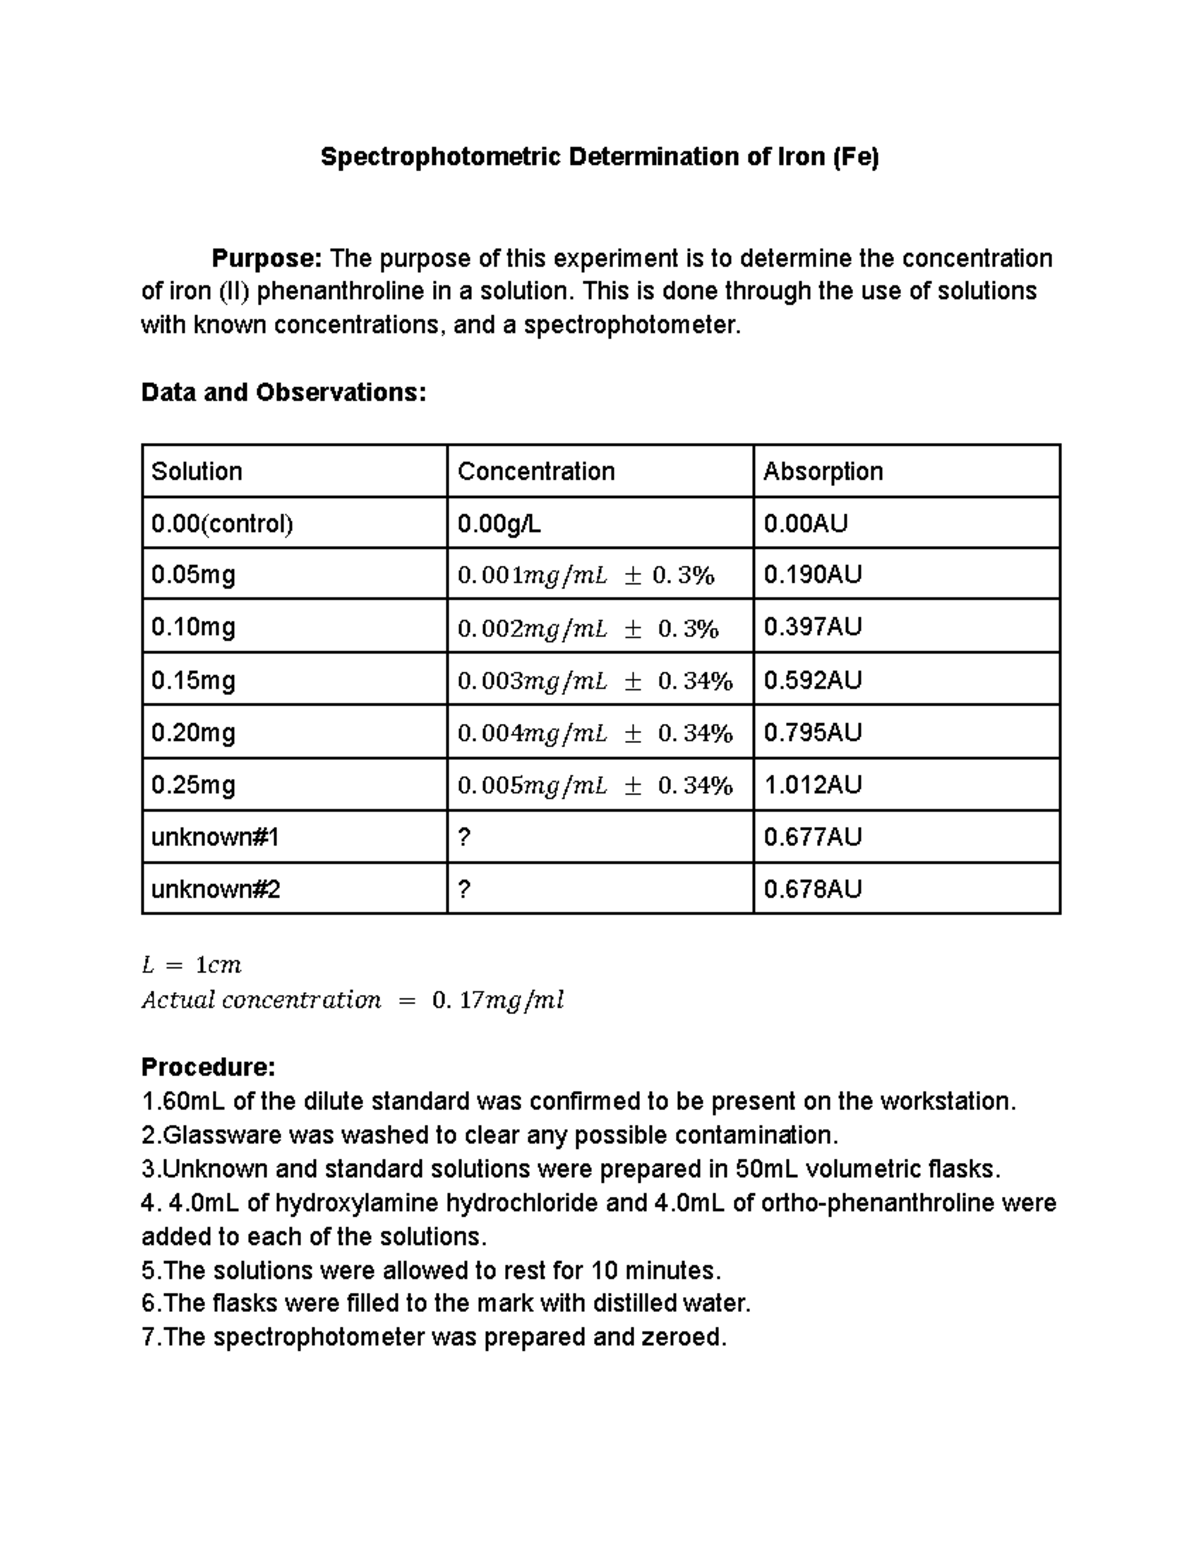
The standard method for determining iron in water samples involves reducing all Fe3+ to Fe2+, complexing with 1,10-phenanthroline, and reading at 510 nm. I have run this procedure hundreds of times across different labs, and the main reason people get poor results usually has nothing to do with the spectrophotometer itself. It is almost always something small done during sample preparation. Here is how the actual workflow goes. You start with your samples and a set of iron standards, typically covering 0 to 10 mg/L depending on your expected concentration. Add hydroxylamine hydrochloride to reduce any Fe3+ to Fe2+. Then add your acetate buffer to bring the pH into the working range, followed by the phenanthroline reagent. Wait thirty minutes for color development, then read absorbance against a reagent blank. Plot your calibration curve and calculate your sample concentrations. The calibration curve should give you an R-squared value above 0.998 if your pipetting is decent. If it is lower than that, your standards are probably contaminated or your cuvettes are scratched. I once spent an entire afternoon trying to figure out why my calibration would consistently drift downward after the third standard. The issue turned out to be a contaminated water bath where I was keeping my samples at room temperature while preparing them. The bath water had trace iron leaching from old fittings. I switched to using freshly deionized water in a separate container and my reproducibility improved immediately. That kind of invisible contamination is the most frustrating part of this work.
Several things that beginners miss about this method are worth noting upfront. The first is that phenanthroline forms a complex with iron that is sensitive to oxidizing agents in your sample. If your water contains chlorine or other oxidants, they will degrade the complex and give you falsely low results. Adding an excess of hydroxylamine hydrochloride helps, but if your oxidant load is high you need to account for it. The second thing is that pH matters more than most protocols suggest. The complex forms optimally between pH 3 and 9, but the most stable readings come in the 3.5 to 4.5 range. Go too high and you get precipitation. Go too low and the color development slows down significantly. Another practical issue is that this method has real limitations. It cannot distinguish between Fe2+ and Fe3+ unless you run separate aliquots with and without the reduction step. That means two sets of measurements just to get total iron versus ferrous iron. It is also vulnerable to interference from heavy metals like copper, lead, and zinc at elevated concentrations, which can form their own complexes with phenanthroline and inflate your reading. Some labs use cyanide or tartrate as masking agents, but handling those adds safety complexity that many teaching labs avoid. When iron concentrations exceed about 10 mg/L, the Beer-Lambert relationship starts to deviate from linearity even though the method is still technically valid. I usually recommend diluting concentrated samples rather than pushing the absorbance past 1.0, since stray light and detector nonlinearity become real factors at higher readings. For very low iron samples below 0.1 mg/L, this method becomes less reliable and you would be better off considering atomic absorption spectroscopy or ICP-OES, which offer lower detection limits without the matrix interferences.
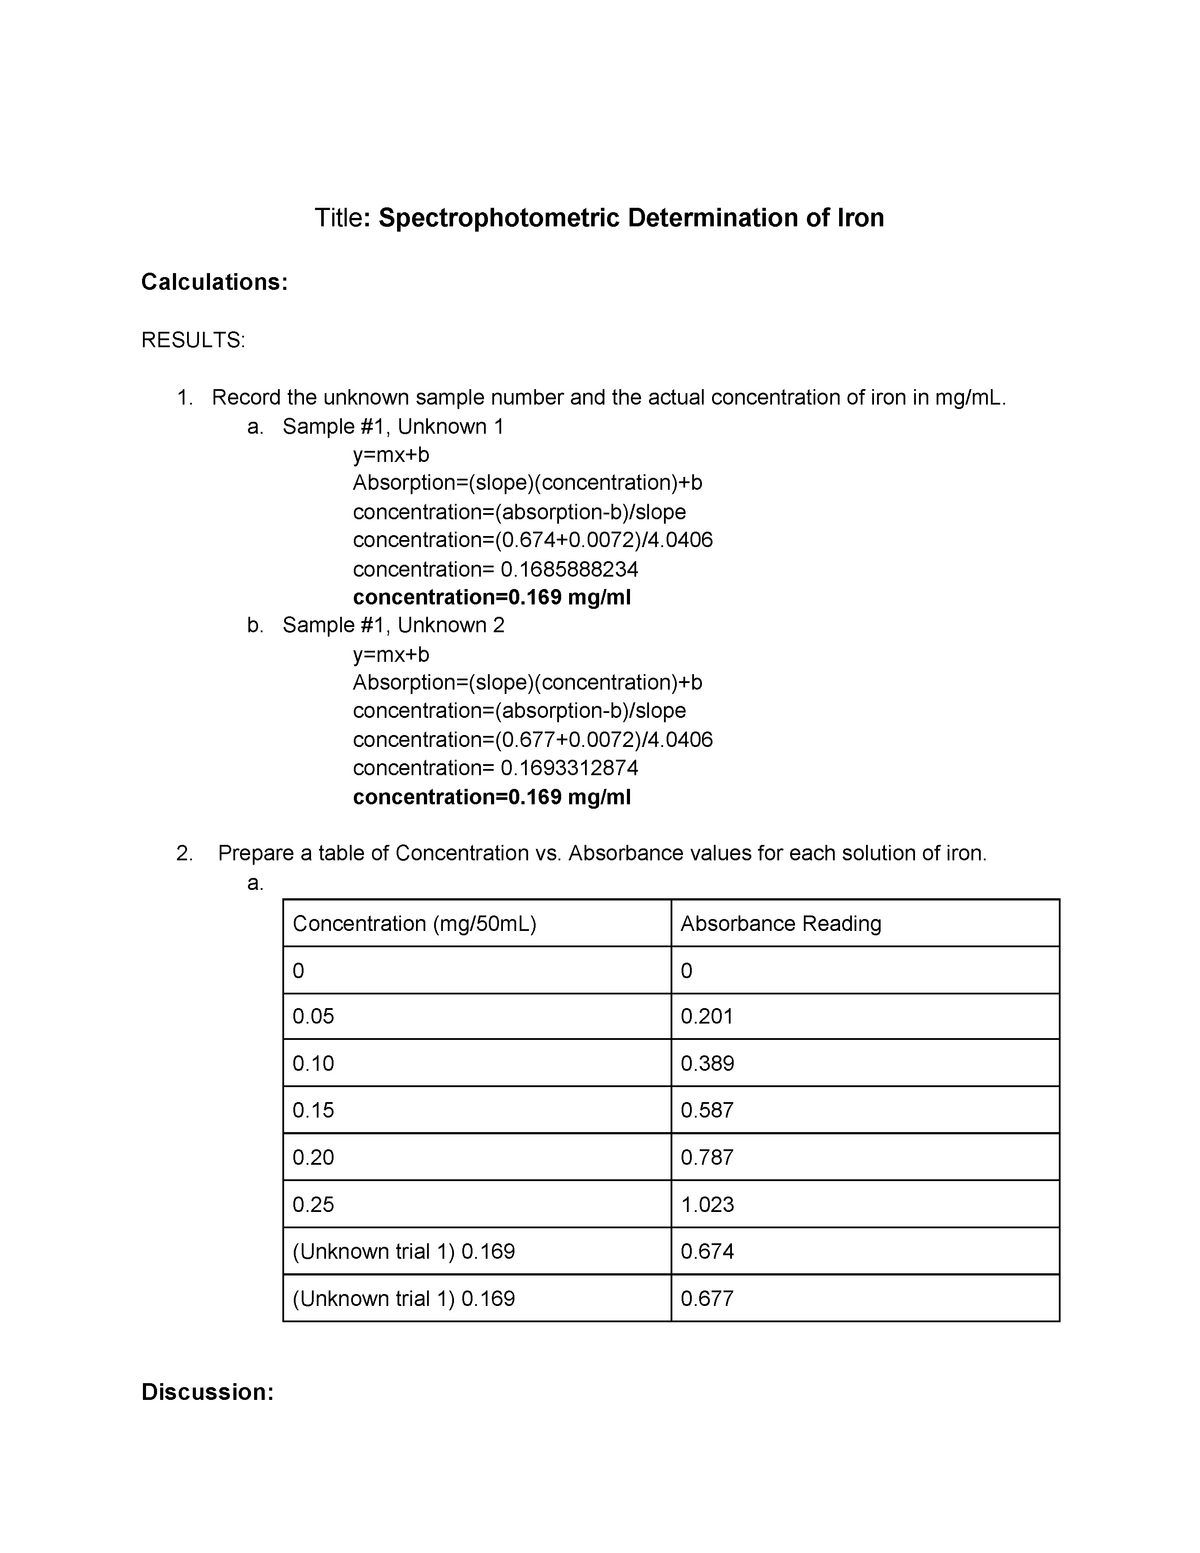
For your lab report, the key sections are your calibration data with the regression equation, your blank correction values, any dilution factors applied to samples, and a discussion of potential interferences specific to your sample matrix. The absorbance readings themselves are straightforward, but showing that you understand where the method falls apart is what separates a competent report from a generic one. Include your uncertainty estimates where possible, especially if you ran replicate measurements. The whole procedure from sample preparation to final reading takes roughly forty-five minutes to an hour for a batch of ten samples. Planning ahead with your standards and samples in order saves significant time compared to working reactively. Keep your reagent bottles capped when not in use since phenanthroline solutions degrade slowly over weeks. Store them in amber glass if you have them, though clear bottles are acceptable for short-term use over a single semester.
Get the Full Details