Working Through Iron Content in Vitamin Tablets
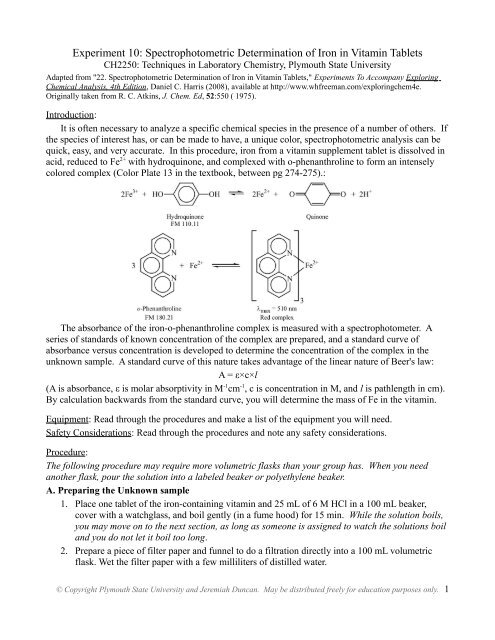
The core calculation for spectrophotometric iron determination in vitamin tablets runs through a Beer-Lambert calibration curve. You prepare standard solutions of known iron concentration, measure their absorbance at 510 nm after complexing with 1,10-phenanthroline, plot absorbance versus concentration, then use the resulting line equation to back-calculate the iron content in your tablet sample. The math itself is straightforward. Getting the chemistry right before you reach the calculator is where most people lose points. Here is the actual sequence I run through when my lab group gets this experiment. First, you crush a known mass of tablets into a fine powder. Accurate weighing matters more than students realize because the final result scales directly with the sample mass you record. A 0.5 mg error on a 0.3 gram sample shifts your answer by nearly two percent, and professors catch that easily. You digest the powder in dilute hydrochloric acid, usually around 1 M, with gentle heating for about fifteen minutes. The iron comes out as Fe3+. The phenanthroline method only complexes with Fe2+, so you need a reducing step. I use hydroxylamine hydrochloride, roughly 10 percent w/v, adding about 1 mL per 25 mL final volume. You let that sit for five minutes at room temperature before adding the phenanthroline reagent. Skip the waiting and your absorbance will be low and inconsistent because the reduction hasn't gone to completion.
The buffer is important too. Acetate buffer brings the pH to roughly 3.5 to 4.5, which is the range where the orange-red Fe(phen)3 2+ complex forms reliably and stays stable. If your pH drifts above 5, you start getting precipitation issues. Below 3 and the complex doesn't form fully. I used to ignore the pH check until I lost an entire batch of standards because I was using tap water instead of deionized water for the buffer. The tap water had alkalinity that pushed the pH to about 7. Nothing precipitated visibly, but the absorbance readings were all over the place. Switched to DI water and the curve went straight through the origin like it should. For the calibration, prepare at least five standard points spanning the expected concentration range. Typical standards run from 1 to 10 micrograms per milliliter of iron. Measure absorbance in a 1 cm cuvette at 510 nm. The regression line should give you an R-squared value above 0.995 if everything is working properly. Below that and you need to check your pipettes, your cuvettes, or whether the spectrophotometer needs warming up longer than the five minutes the manual suggests. I routinely let mine warm for twenty minutes now instead of cutting corners. Once you have your equation, absorbance_sample equals slope times concentration plus intercept, you rearrange to solve for the concentration of your unknown. Then you work backward through the dilution factors. Multiply the calculated concentration by the total volume of your sample solution, divide by the mass of tablet powder used, and multiply by 100 to get percent iron by mass. If your tablet was supposed to contain 65 mg of iron and your 0.3 gram sample was dissolved in 100 mL, the numbers usually land somewhere in that neighborhood. They rarely land exactly on the label value because tablets contain binders and fillers that don't release all their iron into solution, and because manufacturing variability is real.
One thing that trips people up repeatedly is forgetting that the phenanthroline reagent itself adds volume to every solution, including your standards and your samples. Your stock iron solution might be 100 micrograms per milliliter, but once you dilute it with buffer, reducing agent, and phenanthroline, the actual concentration in the cuvette is much lower. Track every dilution step on paper before you start pipetting. I keep a small spreadsheet now instead of trying to do it in my head. Makes errors obvious in a way that mental math never does. The main limitation of this method is interference from other metals. Copper, cobalt, and nickel can also form colored complexes with phenanthroline and throw off your reading. Most vitamin formulations don't have enough of these to matter, but if you are working with a multivitamin that includes significant copper or zinc, you should run a blank containing those matrix components without iron to see if they contribute absorbance at 510 nm. A lot of labs skip this check and just accept the small systematic error. It depends on how precise your instructor expects you to be. Another limitation is that this method measures total soluble iron, not iron in a specific oxidation state or chemical form. If your tablet contains both ferrous and ferric iron, both get measured after the reduction step. That is usually what you want for a general iron content assay, but if you needed speciation data, this approach won't give it to you. Atomic absorption spectroscopy would be the alternative if you need that level of specificity, though it costs more per sample and requires different training.
Get the Full Details
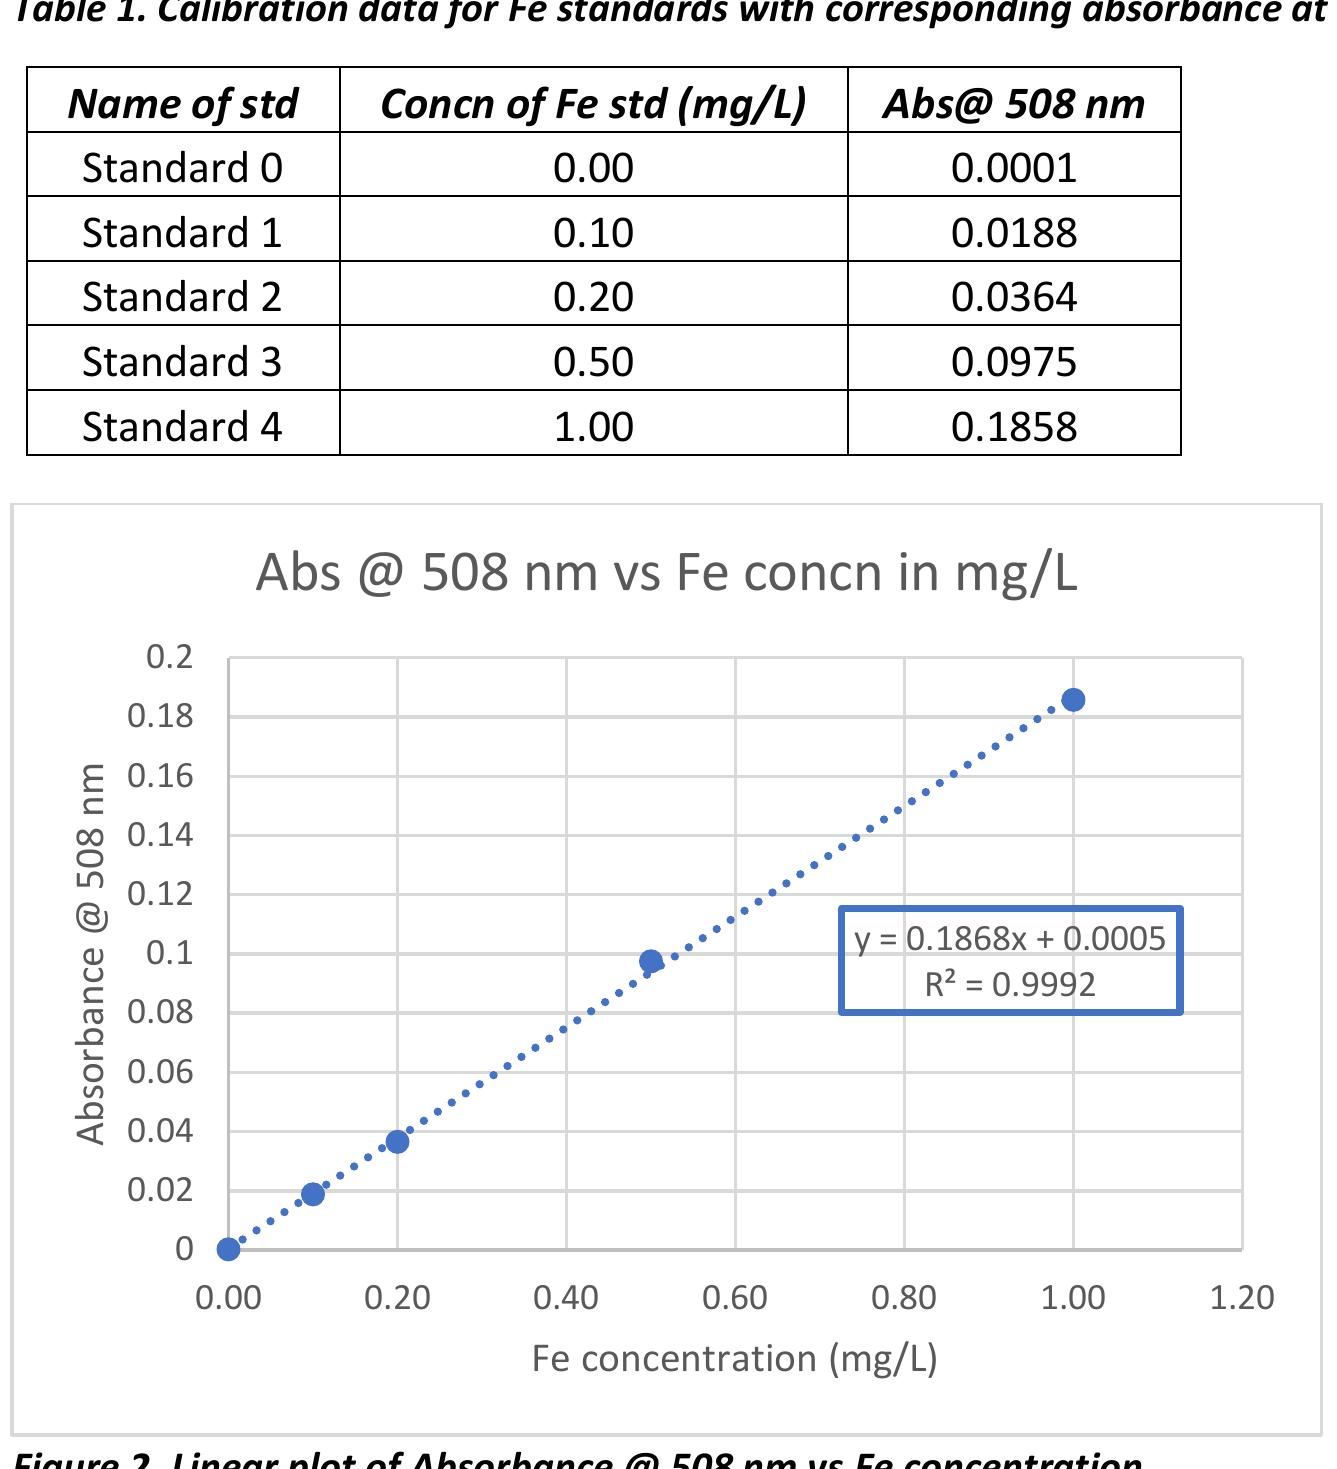
For the final report, include your calibration curve with the equation and R-squared value, your sample absorbance readings, the calculated concentration, all dilution factors shown explicitly, and the final iron content as both milligrams per tablet and percent by mass. Report the standard deviation if you ran replicates. Two replicates minimum, three is better. One reading and a pretty curve is not a valid experiment no matter how clean the data looks.


